Table of Contents
As filed with the Securities and Exchange Commission on September 14, 2026.
Registration No. 333-298617
UNITED STATES
SECURITIES AND EXCHANGE COMMISSION
Washington, D.C. 20549
Amendment No. 1
to
FORM S-1
REGISTRATION STATEMENT
UNDER
THE SECURITIES ACT OF 1933
Electra Therapeutics, Inc.
(Exact name of Registrant as specified in its charter)
| Delaware | 2836 | 83-2193635 | ||
| (State or Other Jurisdiction of Incorporation or Organization) |
Primary Standard Industrial Classification Code Number |
(I.R.S. Employer Identification Number) |
230 E Grand Avenue
Suite S-100
South San Francisco, California 94080
(888) 743-2290
(Address, including zip code, and telephone number, including area code, of Registrant’s principal executive offices)
Quehuong (Kathy) Dong, Pharm.D., M.B.A.
President and Chief Executive Officer
230 E Grand Avenue
Suite S-100
South San Francisco, California 94080
(888) 743-2290
(Name, Address Including Zip Code, and Telephone Number, Including Area Code, of Agent for Service)
Copies to:
| Charles Bair Charles S. Kim Sara Semnani Dylan Kornbluth Cooley LLP 10265 Science Center Drive San Diego, California 92121 (858) 550-6000 |
Chris Clark, C.F.A. Chief Financial Officer Electra Therapeutics, Inc. 230 E Grand Avenue Suite S-100 South San Francisco, California 94080 (888) 743-2290 |
Matthew T. Bush Latham & Watkins LLP 12670 High Bluff Drive San Diego, California 92130 (858) 523-5400 |
Approximate date of commencement of proposed sale to the public: As soon as practicable after this Registration Statement is declared effective.
If any of the securities being registered on this form are to be offered on a delayed or continuous basis pursuant to Rule 415 under the Securities Act of 1933, as amended, check the following box. ☐
If this Form is filed to register additional securities for an offering pursuant to Rule 462(b) under the Securities Act, please check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
If this Form is a post-effective amendment filed pursuant to Rule 462(c) under the Securities Act, check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
If this Form is a post-effective amendment filed pursuant to Rule 462(d) under the Securities Act, check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
Indicate by check mark whether the registrant is a large accelerated filer, an accelerated filer, a non-accelerated filer, a smaller reporting company, or an emerging growth company. See the definitions of “large accelerated filer,” “accelerated filer,” “smaller reporting company,” and “emerging growth company” in Rule 12b-2 of the Exchange Act.
| Large accelerated filer | ☐ | Accelerated filer | ☐ | |||
| Non-accelerated filer | ☒ | Smaller reporting company | ☒ | |||
| Emerging growth company | ☒ | |||||
If an emerging growth company, indicate by check mark if the registrant has elected not to use the extended transition period for complying with any new or revised financial accounting standards provided pursuant to Section 7(a)(2)(B) of the Securities Act. ☐
The Registrant hereby amends this Registration Statement on such date or dates as may be necessary to delay its effective date until the Registrant shall file a further amendment which specifically states that this Registration Statement shall thereafter become effective in accordance with Section 8(a) of the Securities Act of 1933, as amended, or until the Registration Statement shall become effective on such date as the Securities and Exchange Commission, acting pursuant to said Section 8(a), may determine.
Table of Contents
The information in this preliminary prospectus is not complete and may be changed. We may not sell these securities until the registration statement filed with the Securities and Exchange Commission is effective. This preliminary prospectus is not an offer to sell these securities and it is not soliciting an offer to buy these securities in any state or other jurisdiction where the offer or sale is not permitted.
SUBJECT TO COMPLETION, DATED SEPTEMBER 14, 2026
PRELIMINARY PROSPECTUS
21,666,667 Shares
Electra Therapeutics, Inc.
Common stock
We are offering 21,666,667 shares of our common stock. This is our initial public offering, and no public market currently exists for our common stock. We currently expect that the initial public offering price will be between $14.00 and $16.00 per share of our common stock. We have applied to list our common stock on the Nasdaq Global Select Market (Nasdaq) under the symbol “ETRA.” We believe that upon the consummation of this offering, we will meet the standards for listing on Nasdaq, and the consummation of this offering is contingent upon such listing.
We are an “emerging growth company” and a “smaller reporting company” as defined under the U.S. federal securities laws and, as such, may elect to comply with certain reduced public company reporting requirements in future reports after the closing of this offering. See the section titled “Prospectus Summary—Implications of Being an Emerging Growth Company and a Smaller Reporting Company.”
Investing in our common stock involves a high degree of risk. See the section titled “Risk Factors” beginning on page 12 of this prospectus to read about factors you should consider before buying shares of our common stock.
Neither the Securities and Exchange Commission nor any state securities commission has approved or disapproved of these securities or passed upon the accuracy or adequacy of this prospectus. Any representation to the contrary is a criminal offense.
| PER SHARE | TOTAL | |||||||
| Initial public offering price |
$ | $ | ||||||
| Underwriting discounts and commissions (1) |
||||||||
| Proceeds to Electra Therapeutics, Inc., before expenses
|
||||||||
| (1) | See the section titled “Underwriting” for additional information regarding underwriting compensation. |
Delivery of the shares of common stock is expected to be made on or about , 2026.
We have granted the underwriters an option for a period of 30 days from the date of this prospectus to purchase up to an additional 3,250,000 shares of our common stock. If the underwriters exercise the option in full, the total underwriting discounts and commissions payable by us will be $ , and the total proceeds to us, before expenses, will be $ .
| Jefferies | TD Cowen | Evercore ISI | Cantor |
Prospectus dated , 2026
Table of Contents
| PAGE | ||||
| 1 | ||||
| 12 | ||||
| 79 | ||||
| 81 | ||||
| 82 | ||||
| 83 | ||||
| 85 | ||||
| 87 | ||||
| MANAGEMENT’S DISCUSSION AND ANALYSIS OF FINANCIAL CONDITION AND RESULTS OF OPERATIONS |
90 | |||
| 107 | ||||
| 153 | ||||
| 161 | ||||
| 179 | ||||
| 184 | ||||
| 187 | ||||
| 192 | ||||
| MATERIAL U.S. FEDERAL INCOME TAX CONSEQUENCES TO NON-U.S. HOLDERS OF OUR COMMON STOCK |
195 | |||
| 199 | ||||
| 208 | ||||
| 209 | ||||
| 210 | ||||
| F-1 | ||||
Neither we nor the underwriters have authorized anyone to provide you with any information or to make any representations other than those contained in this prospectus, any amendment or supplement to this prospectus or in any free writing prospectus we may authorize to be delivered or made available to you. Neither we nor the underwriters take responsibility for, and can provide no assurance as to the reliability of, any other information that others may give you. We and the underwriters are offering to sell, and seeking offers to buy, shares of our common
i
Table of Contents
stock only in jurisdictions where offers and sales are permitted. The information contained in this prospectus is accurate only as of the date on the front cover of this prospectus, or other earlier date stated in this prospectus, regardless of the time of delivery of this prospectus or any sale of shares of our common stock. Our business, financial condition, results of operations, and prospects may have changed since that date.
For investors outside of the United States: Neither we nor the underwriters have done anything that would permit this offering or possession or distribution of this prospectus in any jurisdiction where action for that purpose is required, other than in the United States. Persons outside the United States who come into possession of this prospectus must inform themselves about, and observe any restrictions relating to, the offering of the shares of our common stock and the distribution of this prospectus outside the United States.
ii
Table of Contents
This summary highlights selected information included elsewhere in this prospectus. This summary does not contain all of the information you should consider before investing in our common stock. You should read this entire prospectus carefully, including the sections titled “Risk Factors,” “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” and “Special Note Regarding Forward-Looking Statements,” and our financial statements and the related notes included elsewhere in this prospectus, before making an investment decision. Unless the context otherwise requires, all references in this prospectus to “Electra Therapeutics,” “Electra,” “Registrant,” “we,” “us,” “our,” and “Company” refer to Electra Therapeutics, Inc.
Overview
We are a late clinical-stage biopharmaceutical company focused on pioneering a new class of precision medicines for the treatment of immune-mediated diseases and cancer. Our novel approach targets signal regulatory proteins (SIRP), a family of cell surface receptors whose expression is restricted to specific immune cell populations, and increases upon activation to enable selective depletion of disease-driving cells while preserving normal immune function. By replacing broad immunosuppression with selective elimination of principal cells that drive disease, we believe our approach can do for immune-mediated diseases what precision oncology has done for cancer, transforming the treatment paradigm for patients. To our knowledge, we are the first company to advance this SIRP-targeted precision immune cell depletion approach into clinical development and demonstrate proof-of-concept in humans.
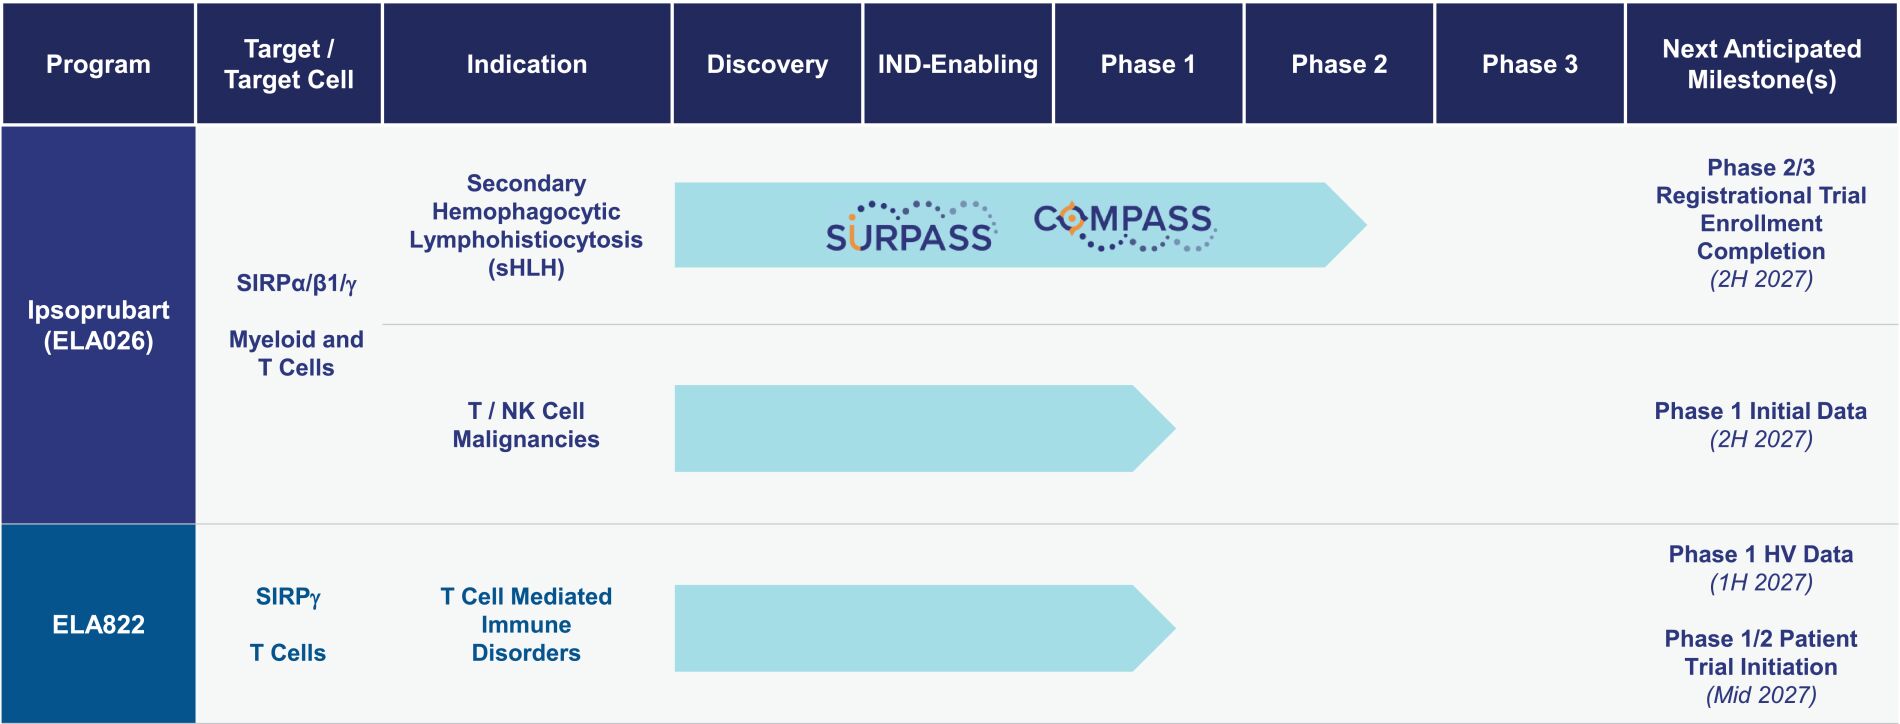
Our lead product candidate is ipsoprubart, a novel pan-SIRP monoclonal antibody designed to selectively deplete pathological myeloid cells and T cells via binding to SIRPa/ß1/g, which is currently in a global registrational program for patients with secondary hemophagocytic lymphohistiocytosis (sHLH). In our Phase 1b trial, ipsoprubart was generally well tolerated and demonstrated a 100% 8-week overall survival (OS) rate and 100% overall response rate (ORR) in 12 frontline patients with malignancy-associated HLH (mHLH), the largest subset of sHLH and the population associated with the poorest outcomes. Of the 12 mHLH frontline patients, 10 achieved a partial response (PR) and two achieved a modified complete response (mCR). We are conducting SURPASS, our global Phase 2/3 registrational trial in newly diagnosed, treatment-naïve sHLH patients, as well as COMPASS, our natural history study designed to provide an external control comparator for SURPASS. We expect to complete enrollment in our registrational program in the second half of 2027. Ipsoprubart has received Breakthrough Therapy designation (BTD) from the U.S. Food and Drug Administration (FDA) and PRIority MEdicine (PRIME) designation from the European Medicines Agency (EMA), each granted for the treatment of sHLH broadly.
In addition, eight evaluable sHLH patients with underlying T or B cell lymphomas treated with ipsoprubart across our Phase 1b trial and Expanded Access Program who had tumor response measurements within weeks of treatment ipsoprubart demonstrated a 100% objective tumor response rate, with seven out of eight patients achieving a complete response (CR). We are conducting a Phase 1 clinical trial evaluating ipsoprubart as a monotherapy in patients with relapsed/refractory T cell and natural killer (NK) cell malignancies, with initial data expected in the second half of 2027.
We are also advancing ELA822, a novel SIRPg-specific monoclonal antibody designed to selectively deplete activated T cells, with potential applications across chronic T cell-mediated immune and inflammatory diseases. In August 2026, we obtained regulatory clearance and initiated a Phase 1 clinical trial of ELA822 in healthy volunteers (HV) in Europe, with data expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027, subject to regulatory approval.
We have generated a proprietary library of SIRP-targeted antibodies with distinct profiles, developed over years of dedicated discovery and engineering. This platform provides substantial flexibility to align antibody design with the cellular biology and therapeutic objectives of each program, and we believe it positions us to pursue a
1
Table of Contents
broad range of indications involving pathogenic SIRP-expressing cells without requiring additional de novo discovery efforts. From our proprietary library of SIRP-targeted antibodies, we have advanced two product candidates, ipsoprubart and ELA822, each designed for distinct disease settings, as shown in the chart below:
Ipsoprubart for the Treatment of Secondary Hemophagocytic Lymphohistiocytosis (sHLH)
Ipsoprubart (formerly known as ELA026), our lead product candidate, is a novel pan-SIRP antibody engineered with an enhanced-effector Fc domain for rapid, potent depletion of pathogenic myeloid cells and T cells. We are developing ipsoprubart initially for sHLH, a severe hyperinflammatory syndrome with no broadly approved therapies and consistently poor survival over the past two decades. In mHLH, which accounts for approximately 45% of all sHLH cases and is associated with the poorest outcomes, historical 2-month OS rates with existing treatments are approximately 50%. Patients with sHLH are typically diagnosed and treated by hematologist-oncologists in both outpatient and inpatient settings. The diagnosed incidence in the United States reached approximately 12,600 in 2025, reflecting an 11.5% compound annual growth rate since 2021, and would reach nearly 17,500 in 2028 assuming a similar annual growth rate. We believe this estimate is conservative as it excludes misdiagnosed, undiagnosed, and non-International Classification of Diseases (ICD) (standardized diagnosis codes used to classify medical conditions) coded patients. As disease awareness increases, underlying triggering conditions become more prevalent, and diagnostic approaches become simpler, we believe the diagnosed incidence will continue to rise.
In our completed Phase 1b trial, ipsoprubart was generally well tolerated and achieved 100% OS at 8 weeks in 12 frontline mHLH patients, alongside a 100% ORR, rapid improvement of key pharmacodynamic (PD) biomarkers, and a 100% hospital discharge rate in all 11 patients evaluable for discharge (one patient withdrew prior to discharge assessment). Additional clinical experience beyond mHLH supports the potential of ipsoprubart as a treatment for sHLH broadly. Based on these data, ipsoprubart has received BTD from the FDA and PRIME designation from the EMA, each granted for the treatment of sHLH broadly. We initiated a global registrational program for ipsoprubart in sHLH consisting of SURPASS, a Phase 2/3 registrational trial, and COMPASS, our natural history study designed to provide an external control comparator for SURPASS, with enrollment completion expected in the second half of 2027.
Ipsoprubart for the Treatment of T/NK Cell Malignancies
In eight patients treated across our Phase 1b trial and Expanded Access Program, who had tumor response measurements within weeks of treatment, ipsoprubart achieved a 100% objective tumor response rate, including an 88% CR rate. One patient with heavily refractory T cell lymphoma treated with ipsoprubart as a monotherapy experienced a CR with respect to both sHLH and the underlying T cell lymphoma. The patient was alive as of the last follow-up, more than one year post treatment. T/NK cell malignancies remain areas of significant unmet medical need, particularly in the relapsed/refractory setting where available therapies demonstrate low CR rates, limited durability, and poor long-term outcomes. We believe these diseases represent
2
Table of Contents
a substantial commercial opportunity and we estimate a diagnosed incidence rate of more than 13,000 annually in the United States across the T/NK cell malignancy subtypes. Based on the anti-tumor activity observed in the patients from the Phase 1b trial and Expanded Access Program, we have initiated a Phase 1 clinical trial of ipsoprubart in relapsed/refractory T/NK cell malignancies and expect initial data in the second half of 2027.
ELA822 for the Treatment of Chronic Immune and Inflammatory Diseases
ELA822, our second product candidate, is a novel SIRPg-specific antibody designed to selectively deplete activated, pathogenic T cells while sparing naïve and regulatory myeloid cells and T cells – a target profile we believe is well suited for a broad range of chronic T cell-mediated immune and inflammatory diseases where durable efficacy and long-term tolerability are important therapeutic objectives. Nonclinical data for ELA822 demonstrated selective depletion of activated T cells and in vivo activity in humanized models of T cell-driven diseases, including giant cell arteritis (GCA) and graft-versus-host disease (GvHD). ELA822 was well tolerated in non-human primate (NHP) studies. We have obtained regulatory clearance and initiated a Phase 1 clinical trial of ELA822 in healthy volunteers in Europe, with data expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027.
Our Approach: Precision SIRP-Targeted Immune Cell Depletion
The Unmet Need in Immune-Mediated Diseases
Pathogenic myeloid cells and T cells are key drivers of immune dysregulation across a broad range of diseases, including acute hyperinflammatory syndromes, chronic inflammatory conditions, and malignancies. Aberrant activation of these cells can lead to excessive release of numerous proinflammatory cytokines and mediators, driving tissue damage, systemic inflammation, and end-organ dysfunction. Current therapies are often limited either by broad immunosuppression associated with non-selective immune modulation or by insufficient disease control resulting from narrowly targeting individual cytokines.
We believe that selectively eliminating activated pathogenic cells, while preserving host defense and immune homeostasis, may provide a more effective therapeutic approach for diseases driven by dysregulated myeloid cell and T cell activity.
SIRP Expression as a Marker of Immune Cell Activation
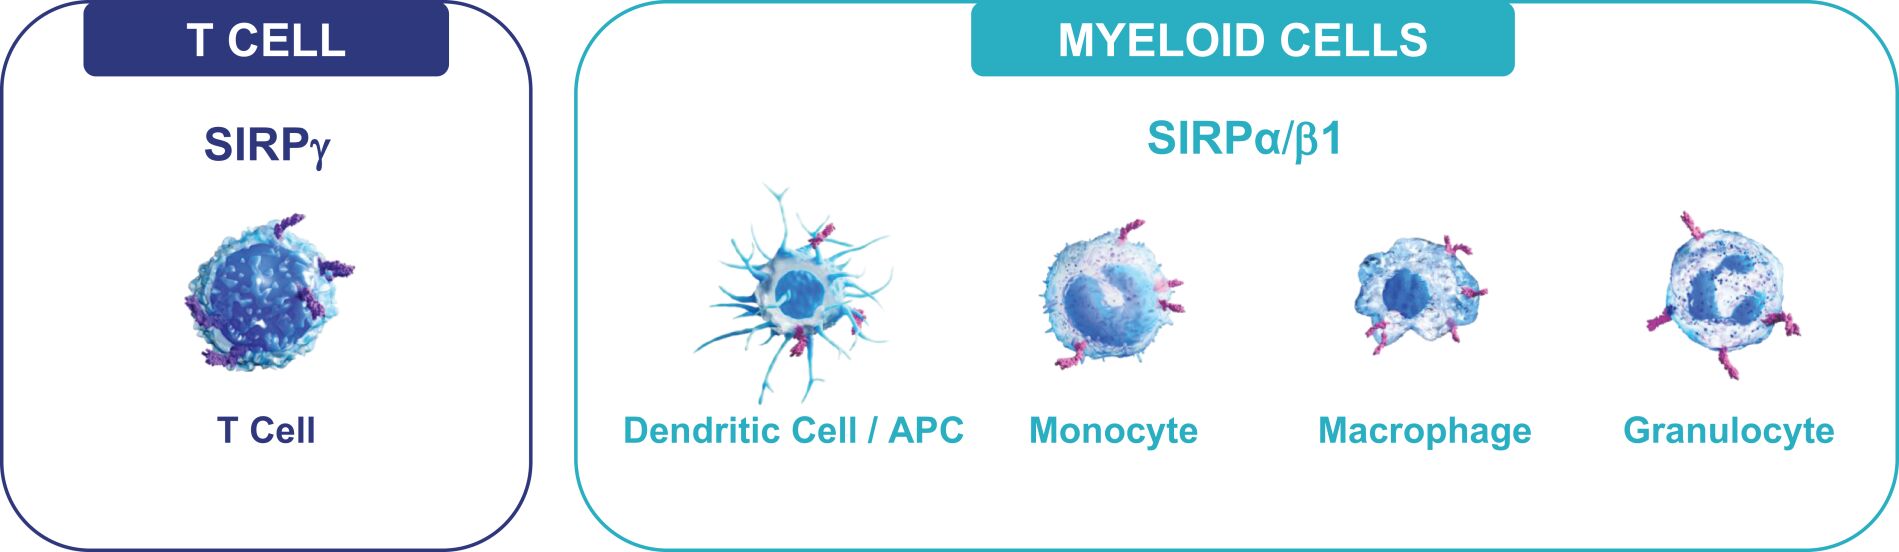
Our approach leverages the unique biology of SIRP to enable selective depletion of activated pathogenic cells. SIRP is a family of cell surface receptors involved in regulating immune homeostasis and inflammatory responses. Their expression is largely restricted to defined immune cell populations and can increase significantly as these cells become activated. This activation-dependent expression pattern creates a molecular marker, or flag, that can be targeted to selectively deplete disease-driving cells while sparing naïve and regulatory immune populations necessary for normal immune function.
The principal members of the SIRP family include SIRPa and SIRPß1, expressed on dendritic cells and antigen-presenting cells (APCs), monocytes, macrophages, and granulocytes; and SIRPg, expressed predominantly on T cells, as shown in the figure below:
3
Table of Contents
Our Distinct Approach to Precision SIRP-Targeted Cell Depletion
Historically, drug development involving SIRP has focused primarily on disrupting the CD47-SIRPa interaction, an innate immune checkpoint commonly referred to as a “don’t eat me” signaling pathway that may be exploited by cancer cells to evade immune surveillance. These approaches have largely been pursued in oncology indications and, to date, have demonstrated limited monotherapy activity and encountered clinical development challenges.
Our approach differs fundamentally from these prior efforts. Rather than disrupting the CD47-SIRPa signaling, we seek to preserve this pathway to maintain immune surveillance while leveraging the lineage-restricted and activation-dependent expression pattern of SIRP to selectively deplete activated pathogenic cells.
Our antibodies are designed to selectively bind SIRP-expressing cells without blocking or otherwise modulating CD47-SIRPa signaling. Our approach is to mediate therapeutic activity through antibody-mediated elimination of the bound cell, achieved through antibody-dependent cellular cytotoxicity (ADCC) and antibody-dependent cellular phagocytosis (ADCP), resulting in elimination of the targeted pathogenic cell. Selectivity is enabled by differential SIRP expression between activated cells and naïve or regulatory immune cell populations. To our knowledge, we are the first company to advance this SIRP-targeted precision immune cell depletion approach into clinical development and demonstrate proof-of-concept in humans.
Our Strengths and Capabilities
We have built differentiated capabilities spanning SIRP-targeted antibody discovery and the underlying SIRP biology, resulting in our current pipeline. Together, we believe the following factors will enable us to achieve our goal of developing, commercializing, and maximizing the impact of precision immune cell depletion therapies across multiple disease areas:
| ∎ | Proprietary and Expansive SIRP-Targeted Antibody Library with Broad Therapeutic Optionality |
| ∎ | Deep Knowledge and Expertise in SIRP Biology |
| ∎ | Proven Track Record of Drug Development and Commercialization in Immunology, Hematology and Oncology |
| ∎ | Disciplined Research and Development Strategy and Efficient Execution for Advancing Novel Therapies |
Our Strategy
Our goal is to become a leading company in precision immune cell depletion with first-in-class or best-in-class SIRP-targeted therapies designed to transform treatment for patients with immune-mediated diseases and cancer. To achieve this goal, we intend to pursue the following:
| ∎ | Advance ipsoprubart through our registrational program as a novel treatment for patients with sHLH |
| ∎ | Expand the therapeutic application of ipsoprubart across hematologic malignancies and other inflammatory indications |
| ∎ | Build a focused and capital-efficient commercial organization, initially centered on ipsoprubart |
| ∎ | Advance ELA822 and demonstrate proof-of-concept in clinical trials for chronic T cell-mediated immune and inflammatory diseases |
| ∎ | Expand our pipeline in SIRP-targeted precision immunology and oncology |
| ∎ | Selectively explore value-creating strategic collaborations to maximize the value of our programs |
Summary of Risk Factors
Investing in our common stock involves significant risks. You should carefully consider the risks described in the section titled “Risk Factors” immediately following this prospectus summary and elsewhere in this prospectus before making a decision to invest in our common stock. If we are unable to successfully address
4
Table of Contents
these risks and challenges, our business, financial condition, results of operations, or prospects could be materially and adversely affected. In such case, the trading price of our common stock would likely decline, and you may lose all or part of your investment. Below is a summary of some of the more significant risks we face.
| ∎ | We have a limited operating history and have no products approved for commercial sale, which may make it difficult for investors to evaluate our business, likelihood of success, and viability. |
| ∎ | We have incurred significant operating losses since our inception and have not generated any revenue. We expect to incur significant losses for the foreseeable future and may never generate any revenue or become profitable, or if we achieve profitability, we may not be able to maintain it. |
| ∎ | Even if this offering is successful, we will require substantial additional capital to finance our operations and achieve our goals. If we are unable to raise capital when needed or on terms acceptable to us, we may be forced to delay, reduce, or eliminate our research or development programs, any future commercialization efforts or other operations. |
| ∎ | We are substantially dependent on the success of our lead product candidate, ipsoprubart, and our other product candidate, ELA822. If we are unable to advance the development of, receive regulatory approval for, and ultimately successfully commercialize ipsoprubart or ELA822, or experience significant delays in doing so, our business will be materially harmed. |
| ∎ | There are currently no approved therapies for the treatment of sHLH broadly, and our research and development activities related to ipsoprubart for the treatment of sHLH broadly may never lead to an approved product. Our discovery and development programs are focused on SIRP-targeted precision immune cell depletion for patients with immune-mediated diseases and cancer, and the scientific discoveries that form the basis for our efforts to discover and develop product candidates are relatively new. |
| ∎ | Drug development is a lengthy and expensive process, the outcome of clinical testing is inherently uncertain, and results of earlier nonclinical studies and clinical trials may not be predictive of future clinical trial results. We may incur additional costs or experience delays in completing, or ultimately be unable to complete, the development and commercialization of ipsoprubart, ELA822 and any future product candidates for many reasons, including a failure to replicate positive results from earlier nonclinical studies or clinical trials in ongoing or future nonclinical studies or clinical trials. |
| ∎ | If we experience delays or difficulties in the enrollment of patients in clinical trials, our receipt of necessary regulatory approvals could be delayed or prevented. |
| ∎ | The incidence and prevalence for target patient populations of one or more of our current product candidates have not been established with precision. If the market opportunities for our current or future product candidates are smaller than we estimate or if any approval that we may obtain is based on a narrower definition of the patient population, our revenue and ability to achieve profitability will be adversely affected, possibly materially. |
| ∎ | Our future performance is dependent on our ability to retain key employees and to attract, retain and motivate qualified personnel and manage our human capital. |
| ∎ | We face substantial competition, which may result in others discovering, developing or commercializing products before or more successfully than we do. |
| ∎ | If we or our future licensors are unable to obtain, maintain, defend, and enforce patent and other intellectual property protection for our current or future product candidates and technologies in the United States or other countries, or if the scope of the patent or other intellectual property protection obtained is not sufficiently broad, we may not be able to compete effectively. |
| ∎ | We cannot ensure that patent rights relating to inventions described and claimed in our pending patent applications will issue or that patents based on our patent applications will not be challenged and rendered invalid and/or unenforceable. |
| ∎ | We are dependent on sole source and limited source suppliers for certain drug products, raw materials, samples, components, and other materials used in our product candidates. If we are unable |
5
Table of Contents
| to source these supplies on a timely basis or establish longer-term contracts with our contract manufacturing organizations (CMOs), we will not be able to complete our clinical trials on time and the development of our product candidates may be delayed. |
| ∎ | We rely, and intend to continue to rely, on third parties to conduct our clinical trials and perform some of our research and potential nonclinical studies. If these third parties do not satisfactorily carry out their contractual duties, fail to comply with applicable regulatory requirements or do not meet expected deadlines, our development programs may be delayed or subject to increased costs or we may be unable to obtain marketing authorization, each of which may have an adverse effect on our business, financial condition, results of operations, and prospects. |
| ∎ | We rely on third parties to manufacture our product candidates and clinical product supplies and we may not be able to obtain adequate supplies at a reasonable cost or in a timely way. |
| ∎ | The regulatory approval process is highly uncertain, and we may be unable to obtain, or may be delayed in obtaining, U.S. or foreign regulatory approval and, as a result, unable to commercialize ipsoprubart, ELA822 and any future product candidates. Even if we believe our current, or planned clinical trials are successful, regulatory authorities may not agree that they provide adequate data on safety or efficacy. |
| ∎ | No public market for our common stock currently exists, and an active and liquid trading market for our common stock may never develop. As a result, you may not be able to resell your shares of common stock at or above the initial public offering price. |
Corporate Information
We were originally incorporated under the laws of the State of Delaware in October 2018 as a wholly-owned subsidiary of Star Therapeutics LLC (Star), a private biotechnology company. In February 2022, in connection with our Series B Preferred Stock financing, we became a majority-owned subsidiary of Star. In June 2023, Star formed Electra Therapeutics LLC (Electra LLC) and contributed its shares of our Series A redeemable convertible preferred stock to Electra LLC in exchange for units of Electra LLC. In connection with being spun out of Star, Star then distributed in-kind the units of Electra LLC to its members, resulting in Star no longer being our parent company or otherwise holding any shares in us (the Spin-Out). As a result of the Spin-Out, we are no longer directly affiliated with Star. Our principal executive office is located at 230 E Grand Avenue, Suite S-100, South San Francisco, California 94080, and our telephone number is (888) 743-2290. Our website is www.electra-therapeutics.com. We have included our website in this prospectus solely as a textual reference. The information contained in, or accessible through, our website is not incorporated by reference into this prospectus, and you should not consider any information contained in, or that can be accessed through, our website as part of this prospectus or in deciding whether to purchase our common stock.
This prospectus contains references to our trademarks and to trademarks, tradenames and service marks belonging to other entities. Solely for convenience, trademarks and trade names referred to in this prospectus, including logos, artwork and other visual displays, may appear without the ® or ™ symbols, but such references are not intended to indicate, in any way, that we will not, or their respective owners will not assert, to the fullest extent under applicable law, our or their rights thereto, as applicable. We do not intend our use or display of other companies’ trade names or trademarks to imply a relationship with, or endorsement or sponsorship of us by, any other companies.
Implications of Being an Emerging Growth Company and a Smaller Reporting Company
We are an “emerging growth company” as defined in the Jumpstart Our Business Startups Act (JOBS Act), and we may remain an emerging growth company for up to five years following the closing of this offering. For so long as we remain an emerging growth company, we are permitted and intend to rely on certain exemptions from various public company reporting requirements, including not being required to have our internal control over financial reporting audited by our independent registered public accounting firm pursuant to Section 404(b) of the Sarbanes-Oxley Act of 2002, as amended (Sarbanes-Oxley Act), reduced disclosure
6
Table of Contents
obligations regarding executive compensation in our periodic reports and proxy statements, and exemptions from the requirements of holding a nonbinding advisory vote on executive compensation and any golden parachute payments not previously approved. In particular, in this prospectus, we have provided only two years of audited financial statements and have not included all of the executive compensation-related information that would be required if we were not an emerging growth company. Accordingly, the information contained herein may be different than the information you receive from other public companies in which you hold stock.
In addition, the JOBS Act provides that an emerging growth company can take advantage of an extended transition period for complying with new or revised accounting standards. This provision allows an emerging growth company to delay the adoption of some accounting standards until those standards would otherwise apply to private companies. We have elected to take advantage of the benefits of this extended transition period and, therefore, we are not subject to the same requirements to adopt new or revised accounting standards as other public companies that are not emerging growth companies; however, we may adopt certain new or revised accounting standards early. We would cease to be an emerging growth company upon the earliest to occur of: (i) the last day of the fiscal year in which we have $1.235 billion or more in annual revenue; (ii) the date on which we first qualify as a large accelerated filer under the rules of the U.S. Securities and Exchange Commission (SEC); (iii) the date on which we have, in any three-year period, issued more than $1.0 billion in non-convertible debt securities; and (iv) the last day of the fiscal year ending after the fifth anniversary of this offering.
We are also a “smaller reporting company” as defined in the Securities Exchange Act of 1934, as amended (Exchange Act). We may continue to be a smaller reporting company even after we are no longer an emerging growth company. We may take advantage of certain of the scaled disclosures available to smaller reporting companies and will be able to take advantage of these scaled disclosures for so long as the market value of our voting and non-voting common stock held by non-affiliates is less than $250.0 million measured on the last business day of our second fiscal quarter, or our annual revenue is less than $100.0 million during the most recently completed fiscal year, and the market value of our voting and non-voting common stock held by non-affiliates is less than $700.0 million measured on the last business day of our second fiscal quarter.
7
Table of Contents
THE OFFERING
| Common stock offered by us |
21,666,667 shares. |
| Option to purchase additional shares of common stock |
We have granted the underwriters an option to purchase up to 3,250,000 additional shares of our common stock from us at the initial public offering price, less the underwriting discounts and commissions, at any time within 30 days of the date of this prospectus. |
| Common stock to be outstanding immediately after this offering |
61,100,288 shares (or 64,350,288 shares if the underwriters exercise their option to purchase additional shares in full). |
| Use of proceeds |
We estimate that the net proceeds to us from the sale of our common stock in this offering will be approximately $296.6 million (or approximately $341.9 million if the underwriters exercise their option to purchase additional shares in full), assuming an initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. |
| We currently intend to use the net proceeds from this offering, together with our existing cash, cash equivalents and marketable securities, (i) to advance the development of ipsoprubart in sHLH through our ongoing Phase 2/3 global registrational trial topline results and to support BLA submission and commercial readiness activities, (ii) to advance the development of ipsoprubart in T/NK cell malignancies through our ongoing Phase 1 clinical trial, (iii) to advance the development of ELA822 through our ongoing Phase 1 clinical trial in healthy volunteers and initiation of a planned Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, and (iv) for other research and development activities, as well as for capital expenditures, working capital and other general corporate purposes. See the section titled “Use of Proceeds” for additional information. |
| Risk factors |
See the section titled “Risk Factors” for additional information and a discussion of factors you should carefully consider before deciding to invest in our common stock. |
| Proposed Nasdaq Global Select Market trading symbol |
“ETRA” |
The number of shares of our common stock to be outstanding after this offering set forth above is based on 39,433,621 shares of our common stock outstanding as of June 30, 2026, after giving effect to the automatic conversion of all outstanding shares of our redeemable convertible preferred stock into 39,404,292 shares of our common stock in connection with the closing of this offering, and excludes:
| ∎ | 5,701,943 shares of our common stock issuable upon the exercise of stock options outstanding under the 2022 Stock Plan (2022 Plan) as of June 30, 2026, with a weighted-average exercise price of $4.92 per share; |
8
Table of Contents
| ∎ | 259,921 shares of our common stock issuable upon the exercise of stock options outstanding granted under the 2022 Plan subsequent to June 30, 2026, with a weighted-average exercise price of $7.03 per share; |
| ∎ | 2,863,183 shares of our common stock issuable upon the exercise of stock options to be granted to certain of our employees, executive officers and non-employee directors under our 2026 Plan, which will become effective upon the execution and delivery of the underwriting agreement for this offering, with an exercise price per share that is equal to the initial public offering price per share in this offering (the IPO Option Grants); |
| ∎ | 13,446,281 shares of our common stock reserved for future issuance under our 2026 Plan (inclusive of the shares issuable under the IPO Option Grants), which will become effective upon the execution and delivery of the underwriting agreement for this offering (which shares include 7,697,039 new shares plus the number of shares (not to exceed 5,749,242 shares) that remain available for the issuance of awards under the 2022 Plan, at the time the 2026 Plan becomes effective, as more fully described in the section titled “Executive and Director Compensation—Equity Benefit Plans”), as well as any automatic increases in the number of shares of our common stock reserved for future issuance under the 2026 Plan; and |
| ∎ | 611,682 shares of our common stock reserved for future issuance under our 2026 Employee Stock Purchase Plan (ESPP), which will become effective upon the execution and delivery of the underwriting agreement for this offering, as well as any annual automatic increases in the number of shares of our common stock reserved for future issuance under the ESPP. |
Unless otherwise indicated, the information in this prospectus reflects and assumes the following:
| ∎ | the automatic conversion, in accordance with our existing certificate of incorporation, of all outstanding shares of our redeemable convertible preferred stock into an aggregate of 39,404,292 shares of our common stock immediately prior to the closing of this offering; |
| ∎ | no exercise of the outstanding stock options described above; |
| ∎ | no exercise by the underwriters of their option to purchase up to 3,250,000 additional shares of our common stock from us in this offering; |
| ∎ | the filing and effectiveness of our amended and restated certificate of incorporation in Delaware in connection with the closing of this offering; and |
| ∎ | a one-for-1.5002 reverse stock split of our common stock and redeemable convertible preferred stock effected on September 10, 2026. |
9
Table of Contents
SUMMARY FINANCIAL DATA
The following tables summarize our financial data as of and for the periods indicated. We have derived the summary statements of operations data for the years ended December 31, 2024 and 2025 from our audited financial statements included elsewhere in this prospectus. We have also derived the summary interim condensed statements of operations data for the six months ended June 30, 2025 and 2026, and the summary interim condensed balance sheet data as of June 30, 2026 from our unaudited interim condensed financial statements included elsewhere in this prospectus. Our audited financial statements and unaudited interim condensed financial statements included elsewhere in this prospectus have been prepared in accordance with U.S. generally accepted accounting principles (U.S. GAAP). Our unaudited interim condensed financial statements were prepared on a basis consistent with our audited financial statements and include, in our opinion, adjustments of a normal and recurring nature that are necessary for the fair statement of the financial information set forth in those statements included elsewhere in this prospectus.
Our historical results presented below are not necessarily indicative of the results to be expected for any future period. The following summaries of our financial data should be read in conjunction with the sections titled “Risk Factors,” “Capitalization,” and “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” and our financial statements and the related notes included elsewhere in this prospectus. The summary financial data included in this section are not intended to replace the financial statements and are qualified in their entirety by our financial statements and the related notes included elsewhere in this prospectus.
| SIX MONTHS ENDED JUNE 30, (UNAUDITED) |
YEAR ENDED DECEMBER 31, |
|||||||||||||||
| (in thousands, except share and per share amounts) | 2025 | 2026 | 2024 | 2025 | ||||||||||||
| Statements of operations data: |
||||||||||||||||
| Operating expenses: |
||||||||||||||||
| Research and development |
$ | 18,792 | $ | 41,637 | $ | 19,267 | $ | 50,387 | ||||||||
| General and administrative |
5,921 | 8,593 | 5,241 | 10,390 | ||||||||||||
| General and administrative—related party |
656 | — | 1,809 | 1,112 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total operating expenses |
25,369 | 50,230 | 26,317 | 61,889 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Loss from operations |
(25,369 | ) | (50,230 | ) | (26,317 | ) | (61,889 | ) | ||||||||
| Other income (expense), net: |
||||||||||||||||
| Interest income |
337 | 1,320 | 1,524 | 1,362 | ||||||||||||
| Interest expense—related party |
(327 | ) | — | — | (1,433 | ) | ||||||||||
| Other income (expense) |
(2 | ) | (2 | ) | 7 | (15 | ) | |||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total other income (expense), net |
8 | 1,318 | 1,531 | (86 | ) | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Net loss |
$ | (25,361 | ) | $ | (48,912 | ) | $ | (24,786 | ) | $ | (61,975 | ) | ||||
|
|
|
|
|
|
|
|
|
|||||||||
| Deemed dividend to Series C redeemable convertible preferred stockholders |
— | (9,675 | ) | — | — | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Net loss attributable to common stockholders |
$ | (25,361 | ) | $ | (58,587 | ) | $ | (24,786 | ) | $ | (61,975 | ) | ||||
|
|
|
|
|
|
|
|
|
|||||||||
| Comprehensive Loss: |
||||||||||||||||
| Net loss |
$ | (25,361 | ) | $ | (48,912 | ) | $ | (24,786 | ) | $ | (61,975 | ) | ||||
| Unrealized gain (loss) on marketable securities |
(1 | ) | (34 | ) | — | 20 | ||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total comprehensive loss |
$ | (25,362 | ) | $ | (48,946 | ) | $ | (24,786 | ) | $ | (61,955 | ) | ||||
|
|
|
|
|
|
|
|
|
|||||||||
| Net loss per share attributable to common stockholders, basic and diluted (1) |
$ | (864.71 | ) | $ | (1,997.58 | ) | $ | (845.10 | ) | $ | (2,113.10 | ) | ||||
| Weighted-average common shares outstanding, basic and diluted (1) |
29,329 | |
29,329 |
|
|
29,329 |
|
|
29,329 |
| ||||||
| Pro forma net loss per share attributable to common stockholders, basic and diluted (unaudited) (2) |
$ | (1.49 | ) | $ | (1.80 | ) | ||||||||||
| Pro forma weighted-average common shares outstanding, basic and diluted (unaudited) (2) |
39,433,621 | 34,515,144 | ||||||||||||||
10
Table of Contents
| (1) | See Note 14 to our audited financial statements and Note 13 to our unaudited interim condensed financial statements included elsewhere in this prospectus for details on the calculation of net loss per share attributable to common stockholders, basic and diluted. |
| (2) | Pro forma net loss per share attributable to common stockholders, basic and diluted, is calculated giving effect to the automatic conversion of all outstanding shares of our redeemable convertible preferred stock into shares of our common stock. Pro forma net loss per share attributable to common stockholders does not include the shares expected to be sold and related proceeds to be received in this offering. Pro forma net loss per share attributable to common stockholders (unaudited) for the year ended December 31, 2025 and for the six months ended June 30, 2026 was calculated using the weighted-average number of shares of our common stock outstanding, including the pro forma effect of the automatic conversion of all outstanding shares of our redeemable convertible preferred stock into shares of our common stock, as if such conversions had occurred at the beginning of the period. |
| AS OF JUNE 30, 2026 | ||||||||||||
| (in thousands) | ACTUAL | PRO FORMA (1) | PRO FORMA AS ADJUSTED (2)(3) |
|||||||||
| Balance sheet data: |
||||||||||||
| Cash, cash equivalents and marketable securities |
$ | 97,725 | $ | 97,725 | $ | 394,325 | ||||||
| Working capital (4) |
86,735 | 86,735 | 383,335 | |||||||||
| Total assets |
106,498 | 106,498 | 403,098 | |||||||||
| Total liabilities … |
14,782 | 14,782 | 14,782 | |||||||||
| Redeemable convertible preferred stock |
311,171 | — | — | |||||||||
| Accumulated deficit |
(221,890 | ) | (221,890 | ) | (221,890 | ) | ||||||
| Total stockholders’ equity (deficit) |
(219,455 | ) | 91,716 | 388,316 | ||||||||
| (1) | Gives effect to the automatic conversion of all outstanding shares of redeemable convertible preferred stock into an aggregate of 39,404,292 shares of our common stock and the related reclassification of the redeemable convertible preferred stock to permanent equity in connection with the closing of this offering. |
| (2) | Gives effect to (i) the pro forma adjustments set forth in footnote (1) above and (ii) our sale of 21,666,667 shares of our common stock in this offering at an assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. |
| (3) | Pro forma as adjusted balance sheet data is illustrative only and will change based on the actual initial public offering price and other terms of this offering determined at pricing. Each $1.00 increase (decrease) in the assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, would increase (decrease) each of our pro forma as adjusted cash, cash equivalents and marketable securities, working capital, total assets and total stockholders’ equity by approximately $20.2 million, assuming that the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same, and after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. We may also increase or decrease the number of shares we are offering. Each one million share increase (decrease) in the number of shares offered by us would increase (decrease) each of our pro forma as adjusted cash, cash equivalents and marketable securities, working capital, total assets and total stockholders’ equity by approximately $14.0 million, assuming that the assumed initial offering price to the public remains the same, and after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. |
| (4) | We define working capital as current assets less current liabilities. See our unaudited interim condensed financial statements and related notes included elsewhere in this prospectus for further details regarding our current assets and current liabilities. |
11
Table of Contents
Investing in our common stock involves a high degree of risk. Before making your decision to invest in shares of our common stock, you should carefully consider the risks described below, together with the other information contained in this prospectus, including in the section titled “Management’s Discussion and Analysis of Financial Condition and Results of Operations” and in our financial statements and the related notes included elsewhere in this prospectus. The risks and uncertainties described below are not the only ones we face. Additional risks and uncertainties that we are unaware of or that we deem immaterial may also become important factors that adversely affect our business. We cannot assure you that any of the events discussed below will not occur. These events could have a material and adverse impact on our business, financial condition, results of operations, and prospects. If that were to happen, the trading price of our common stock could decline, and you could lose all or part of your investment.
Risks Related to Our Limited Operating History, Financial Position and Need for Additional Capital
We have a limited operating history and have no products approved for commercial sale, which may make it difficult for investors to evaluate our business, likelihood of success, and viability.
We are a late clinical-stage biopharmaceutical company with a limited operating history. We were formed in 2018, have no products approved for commercial sale and have never generated any revenue. Drug development is a highly speculative undertaking and involves a substantial degree of risk. It entails substantial upfront capital expenditures and significant risk that any product candidate will fail to demonstrate adequate efficacy or an acceptable safety profile, gain regulatory approval or become commercially viable. Since our inception, we have devoted substantially all of our resources to organizing and staffing our company, business planning, identifying product candidates, establishing our intellectual property portfolio, conducting research and nonclinical studies, including Investigational New Drug Application (IND)-enabling studies, initiating and conducting clinical trials, establishing arrangements with third parties for the manufacture of our product candidates, and providing general and administrative support for these operations.
To date, we have financed our operations primarily through the issuance and sale of shares of our redeemable convertible preferred stock and the issuance of convertible promissory notes. As of June 30, 2026, issuances and sales of our redeemable convertible preferred stock and convertible promissory notes have resulted in aggregate gross proceeds of $301.4 million.
We have not yet demonstrated an ability to successfully obtain regulatory approvals, manufacture a commercial-scale product or arrange for a third party to do so on our behalf, or conduct sales and marketing activities necessary for successful product commercialization. As a result, it may be more difficult for you to accurately predict our likelihood of success and viability than it could be if we had a longer operating history.
In addition, we may encounter unforeseen expenses, difficulties, complications, delays, and other known and unknown factors and risks frequently experienced by clinical-stage biopharmaceutical companies developing targeted product candidates for immune-mediated diseases and cancer. We also may need to transition from a company with a research and development focus to a company capable of supporting commercial activities. We have not yet demonstrated an ability to successfully overcome such risks and difficulties, or to make such a transition. We may not be successful in such a transition. If we do not adequately address these risks and difficulties or successfully make such a transition, our business will suffer.
We have incurred significant operating losses since our inception and have not generated any revenue. We expect to incur significant losses for the foreseeable future and may never generate any revenue or become profitable, or if we achieve profitability, we may not be able to maintain it.
We have incurred significant operating losses in each reporting period since our inception, have not generated any revenue to date and have financed our operations principally through private placements of our redeemable convertible preferred stock and convertible promissory notes. For the years ended December 31, 2024 and 2025, we reported a net loss of $24.8 million and $62.0 million, respectively. For the six months ended June 30, 2025 and 2026, we reported a net loss of $25.4 million and $48.9 million, respectively. As of June 30, 2026, we had an
12
Table of Contents
accumulated deficit of $221.9 million. There is no assurance that we will obtain financing from other sources, or that we will be able to obtain such financing on favorable terms, if at all. Substantially all of our losses have resulted from expenses incurred in connection with the research and development of ipsoprubart and ELA822, and from general and administrative costs associated with our operations. We expect to incur increasing levels of operating losses for the foreseeable future, particularly as we advance ipsoprubart and ELA822 through clinical development and otherwise advance our earlier-stage discovery programs. Our prior losses have had, and combined with expected future losses will continue to have, an adverse effect on our stockholders’ equity and working capital. We expect our research and development expenses to significantly increase in connection with our ongoing and planned nonclinical studies and clinical trials for ipsoprubart and ELA822, as applicable. In addition, our expenses could increase if we are required by the FDA, or any comparable foreign regulatory authority, to perform clinical trials in addition to those currently expected, or if there are any delays in completing our clinical trials or in the nonclinical or manufacturing-related activities associated with the development of our product candidates. Moreover, if we obtain regulatory approval for ipsoprubart and ELA822 or any future product candidates, we will incur significant sales, marketing, manufacturing and distribution expenses in connection with the commercialization of ipsoprubart and ELA822 or any other future product candidates. We may never succeed in these activities and, even if we do, we may never generate any revenue or revenue that is significant enough to achieve profitability.
As a result, we expect to continue to incur significant and increasing net losses for the foreseeable future. Because of the numerous risks and uncertainties associated with developing therapeutic products, we are unable to predict the extent of any future losses or when we will become profitable, if at all. To become and remain profitable, we must succeed in a range of challenging activities, including discovering, developing, completing clinical trials, obtaining regulatory approvals for, and eventually commercializing products that generate significant revenue. We are only in the preliminary stages of a few of these activities.
Even if we do become profitable, we may not be able to sustain or increase our profitability on a quarterly or annual basis. In addition, we expect our financial condition and operating results to fluctuate significantly from quarter-to-quarter and year-to-year due to a variety of factors, many of which are beyond our control. Accordingly, you should not rely on the results of any quarterly or annual periods as indications of future operating performance. If we fail to become and remain profitable, there may be an adverse effect on the value of our company which could impair our ability to raise capital, expand our business, maintain our research and development efforts, diversify our product candidate pipeline, achieve our strategic objectives or even continue our operations. A decline in the value of our company could also cause you to lose all or part of your investment.
Even if this offering is successful, we will require substantial additional capital to finance our operations and achieve our goals. If we are unable to raise capital when needed or on terms acceptable to us, we may be forced to delay, reduce, or eliminate our research or development programs, any future commercialization efforts or other operations.
Developing therapeutic products, including conducting nonclinical studies and clinical trials, is a very time-consuming, expensive and uncertain process that takes years to complete. Our operations have consumed substantial amounts of cash since inception, and we expect our expenses to increase substantially in connection with our ongoing activities, particularly as we advance our lead product candidate, ipsoprubart, and our other product candidate, ELA822, and otherwise advance our earlier-stage discovery programs through clinical development. We expect increased expenses as we continue our research and development, continue our ongoing clinical trials, initiate additional clinical trials, seek to expand our product pipeline and clinical applications, seek regulatory approval for our current and future product candidates, and invest in our organization. In addition, if we obtain regulatory approval for any of our product candidates, we expect to incur significant commercialization expenses related to product manufacturing, marketing, sales and distribution. Furthermore, upon the closing of this offering, we expect to incur additional costs associated with operating as a public company that we did not incur as a private company. Accordingly, we will need to obtain substantial additional funding in connection with our continuing operations.
We had $97.7 million in cash, cash equivalents and marketable securities as of June 30, 2026. Based on our current operating plan, we believe that the net proceeds from this offering, together with our existing cash, cash equivalents and marketable securities will be sufficient for us to fund our operations into 2029. We have based this estimate on assumptions that may prove to be wrong, and we could use our capital resources sooner than we currently expect. Changes beyond our control may occur that would cause us to use our available capital before that
13
Table of Contents
time, including changes in and progress of our drug development activities and changes in regulation. Our future capital requirements will be dependent on many factors, including:
| ∎ | the progress, timing and results of clinical trials and nonclinical studies for ipsoprubart and ELA822, respectively, or any future product candidates; |
| ∎ | further development of our proprietary library of SIRP-targeted antibodies; |
| ∎ | the extent to which we develop, in-license, out-license or acquire any future product candidates or technologies; |
| ∎ | the number and development requirements of any future product candidates that we may pursue, and other indications for our current product candidates that we may pursue; |
| ∎ | the costs, timing and outcome of obtaining regulatory approvals of our current or future product candidates; |
| ∎ | the scope and costs of making arrangements with third-party manufacturers, or establishing manufacturing capabilities, for both clinical and commercial supplies of our current or future product candidates; |
| ∎ | the costs involved in growing our organization to the size needed to allow for the research, development and potential commercialization of our current or future product candidates; |
| ∎ | the costs associated with commercializing any approved product candidates, including establishing sales, marketing, market access, and distribution capabilities; |
| ∎ | to the extent we pursue strategic collaborations, including collaborations to commercialize ipsoprubart, ELA822 or any of our future product candidates, our ability to establish and maintain collaborations on favorable terms, if at all, as well as the timing and amount of any milestone or royalty payments we are required to make or are eligible to receive under such collaborations or our current licenses and platform service agreements; |
| ∎ | the costs associated with completing any post-marketing studies or trials required by the FDA, or other comparable foreign regulatory authorities; |
| ∎ | the revenue, if any, received from commercial sales of ipsoprubart and ELA822 or any of our future product candidates, if any are approved; |
| ∎ | the costs of preparing, filing, and prosecuting patent applications, maintaining and enforcing our intellectual property rights and defending intellectual property-related claims that we may become subject to, including any litigation costs and the outcome of such litigation; and |
| ∎ | the costs associated with potential product liability claims, including the costs associated with obtaining insurance against such claims and with defending against such claims. |
Even if this offering is successful, we will require additional capital to complete our planned nonclinical studies and clinical trials for our current product candidates to obtain regulatory approval, and we anticipate needing to raise additional capital to complete the development of, and eventually commercialize, our product candidates, if approved. Adequate additional financing may not be available to us on favorable terms, or at all. Our ability to raise additional funds will be dependent on financial, economic and market conditions, geopolitical issues and other factors, over which we may have limited or no control. In addition, we may seek additional capital due to favorable market conditions or strategic considerations even if we believe we have sufficient funds for our current or future operating plans. If adequate funds are not available on commercially acceptable terms when needed, we may be forced to delay, reduce or terminate the development or commercialization, if approved, of all or part of our research programs or product candidates or we may be unable to take advantage of future business opportunities, including pursuing new in-licenses and acquisitions. Furthermore, any additional capital-raising efforts may divert our management from their day-to-day activities, which may adversely affect our ability to develop and commercialize our current and any future product candidates, if approved. Changing circumstances, some of which may be beyond our control, could cause us to consume capital significantly faster than we currently anticipate, and we may need to seek additional funds sooner than planned.
We will be required to obtain further funding through public or private equity financings, debt financings, collaboration agreements, licensing arrangements, or other sources of financing, which may dilute our stockholders or restrict our operating activities. We do not have any committed external source of funds. To the extent that we raise additional capital through the sale of equity or convertible debt securities, each investor’s ownership interests
14
Table of Contents
will be diluted, and the terms may include liquidation or other preferences that adversely affect each investor’s rights as a stockholder. Debt financing or preferred equity financing, if available, may involve agreements that include covenants limiting or restricting our ability to take specific actions, such as incurring additional debt, making capital expenditures or declaring dividends. Such restrictions could adversely impact our ability to conduct our operations and execute our business plan. If we raise additional funds through upfront payments or milestone payments pursuant to strategic collaborations with third parties, we may have to relinquish valuable rights to our product candidates or grant licenses on terms that are not favorable to us.
Our failure to raise capital as and when needed or on acceptable terms could significantly harm our business, financial condition, results of operations, and prospects and cause the price of our common stock to decline, and we may have to delay, reduce the scope of, suspend, or eliminate one or more of our research or drug development programs, nonclinical studies, clinical trials, or future commercialization efforts.
The report of our independent registered public accounting firm included a “going concern” explanatory paragraph.
We have concluded that there is substantial doubt about our ability to continue as a going concern, and the report of our independent registered public accounting firm on our financial statements as of and for the year ended December 31, 2025 included an explanatory paragraph indicating that there was substantial doubt about our ability to continue as a going concern. If we are unable to raise additional capital as and when needed, our business, financial condition and results of operations will be materially and adversely affected, and we may be forced to delay our development efforts, limit our activities and reduce research and development costs. If we are unable to continue as a going concern, we may have to liquidate our assets, and the values we receive for our assets in liquidation or dissolution could be significantly lower than the values reflected in our financial statements. The inclusion of a going concern explanatory paragraph by our independent registered public accounting firm, our limited cash resources and our potential inability to continue as a going concern may materially adversely affect our ability to raise new capital, enter into licensing and collaboration arrangements or other contractual relationships with third parties and otherwise execute our development strategy.
Risks Related to Research, Discovery, Development, Regulatory Approval, and Commercialization of Our Product Candidates
We are substantially dependent on the success of our lead product candidate, ipsoprubart, and our other product candidate, ELA822. If we are unable to advance the development of, receive regulatory approval for, and ultimately successfully commercialize ipsoprubart or ELA822, or experience significant delays in doing so, our business will be materially harmed.
We have invested, and plan to continue to invest, a significant portion of our efforts and financial resources in the development of our lead product candidate, ipsoprubart. Our future success is highly dependent on our ability to timely complete successful clinical trials, obtain regulatory approval for, and then successfully commercialize ipsoprubart and our other product candidate, ELA822, which may never occur. We are currently conducting a Phase 2/3 registrational trial of ipsoprubart in sHLH, a Phase 1 clinical trial of ipsoprubart in T cell malignancies and initiated a Phase 1 clinical trial for ELA822 in Europe as our clinical trial application (CTA) for ELA822 was approved in August 2026. Our other potential product candidates are also in earlier stages of development. We currently have no products that are approved for sale in any jurisdiction. We have invested substantially all of our efforts and financial resources in ipsoprubart and ELA822 and conducting nonclinical studies and clinical trials. There can be no assurance that ipsoprubart or ELA822 or any future product candidates we develop will achieve success in their nonclinical studies, clinical trials, or obtain regulatory approval. In the future, we may also become dependent on other product candidates that we may develop or acquire; however, given the stage of our development, approval for commercialization may require additional time and cannot be assured.
Our ability to generate product revenue, which may not occur for many years, if ever, will be heavily dependent on the successful development and eventual commercialization of our lead product candidate, ipsoprubart, our other product candidate, ELA822, and the development of additional product candidates. The success of ipsoprubart, ELA822 and any additional product candidates will be dependent on several factors, including the following:
| ∎ | timely completion of nonclinical studies; |
15
Table of Contents
| ∎ | acceptance of, or allowance to proceed under, any INDs by the FDA, or under comparable CTAs by comparable foreign regulatory authorities for our clinical trials; |
| ∎ | timely and successful enrollment of patients in, and timely and successful completion of, clinical trials with favorable results; |
| ∎ | our ability to enroll adequate participants to allow the results to be generalizable to the U.S. population; |
| ∎ | the frequency and severity of adverse events in clinical trials; |
| ∎ | approval of Biologics License Applications (BLAs) by the FDA or similar foreign regulatory approvals, including the completion of any required post-marketing studies or trials and available funding to perform any post-marketing commitments; |
| ∎ | raising additional funds necessary to complete clinical development of and commercialize our current or future product candidates; |
| ∎ | obtaining, maintaining, expanding and protecting our patent, trade secret and other intellectual property and any regulatory exclusivity for our current and future product candidates; |
| ∎ | making arrangements with third-party manufacturers, or establishing manufacturing capabilities, for both clinical and commercial supplies of our current and future product candidates and ensuring a resilient, effective supply chain that produces supply that outpaces demand; |
| ∎ | developing and implementing marketing and reimbursement strategies, and creating adequate demand forecasts for supply and sales planning; |
| ∎ | establishing sales, marketing and distribution capabilities and launching commercial sales of our product candidates, if and when approved, whether alone or in collaboration with others in a market where promotional sales approaches are rapidly moving to digital platforms; |
| ∎ | demonstration of safety, purity and potency (or efficacy), and acceptable risk-benefit profiles of our product candidates to the satisfaction of the FDA and comparable foreign regulatory authorities; |
| ∎ | acceptance of our product candidates, if and when approved, by patients, the medical community and third-party payors underpinned by adequate health economic data and a meaningful value proposition; |
| ∎ | effectively competing with existing and future therapies; |
| ∎ | obtaining and maintaining third-party payor coverage and adequate reimbursement in both public and private payor spaces; |
| ∎ | obtaining appropriate support from patient advocacy organizations; |
| ∎ | addressing any delays in our clinical trials; and |
| ∎ | maintaining a continued acceptable safety profile of our product candidates following any approval. |
Many of these factors are beyond our control, and it is possible that none of our product candidates will ever obtain regulatory approval even if we expend substantial time and resources seeking such approval. If we do not achieve one or more of these factors in a timely manner or at all, we could experience significant delays or an inability to successfully commercialize our product candidates, which would materially harm our business. For example, our business could be harmed if results of our ongoing clinical trials of ipsoprubart show unexpected adverse events or a lack of efficacy in the patient populations we intend to treat, do not meet the clinical endpoints or if we experience other regulatory or developmental issues.
There are currently no approved therapies for the treatment of sHLH broadly, and our research and development activities related to ipsoprubart for the treatment of sHLH broadly may never lead to an approved product. Our discovery and development programs are focused on SIRP-targeted precision immune cell depletion for patients with immune-mediated diseases and cancer, and the scientific discoveries that form the basis for our efforts to discover and develop product candidates are relatively new.
We are evaluating ipsoprubart to treat sHLH broadly. There are currently no approved therapies for the treatment of sHLH broadly. Our SIRP-targeted precision immune cell depletion approach is novel and scientifically uncertain, and our research and development activities may never lead to approved or marketable products. The discovery and development of precision medicines for patients with immune-mediated diseases and cancer is an emerging field, and the scientific discoveries that form the basis for our efforts to discover and develop product candidates are relatively new. The scientific evidence to support the feasibility of developing product candidates based on these
16
Table of Contents
discoveries is both preliminary and limited. Although we believe, based on our nonclinical work and early clinical trial results, that the depletion of SIRP-expressing immune cells by antibodies via binding to SIRPa/ß1/g will enable selective depletion of disease-driving cells while preserving normal immune function, future clinical results may not confirm this hypothesis or may only confirm it for certain SIRP-expressing immune cells or in certain immune-mediated diseases and cancer.
We cannot be certain that our approach will lead to the development of an approvable or marketable product. We may not succeed in demonstrating safety and efficacy of ipsoprubart for the treatment of sHLH broadly in our ongoing or anticipated clinical trials or in larger-scale clinical trials. Advancing ipsoprubart in development creates significant challenges for us, including:
| ∎ | obtaining regulatory approval, as the FDA and other comparable foreign regulatory authorities have yet to approve a therapy for the treatment of sHLH broadly; |
| ∎ | if ipsoprubart is approved, educating medical personnel regarding the potential efficacy and safety benefit, as well as the challenges, of incorporating our product into their clinical practice; and |
| ∎ | establishing the sales and marketing capabilities upon obtaining any regulatory approvals to gain market acceptance. |
Our scientific approach to target cell surface receptors whose expression is restricted to specific immune cell populations and increases upon activation to selectively deplete disease-driving cells while preserving normal immune function may not be successful in addressing the limitations of broad immunosuppression or creating product candidates that ultimately succeed in development, regulatory approval or commercialization. We may also be unsuccessful in using our approach to identify additional product candidates, and any of our product candidates may be shown to have harmful side effects or other characteristics that necessitate additional clinical testing or make them unmarketable or unlikely to receive or maintain regulatory approval. In particular, any failure of one of our development programs (or a competitor’s product or program pursuing a similar approach) could create a perception that our other programs are less likely to succeed or that our overall approach is not viable.
We believe we have had favorable nonclinical study and early clinical trial results; however, we are early in our development efforts, including with respect to ipsoprubart and ELA822, and may not succeed in demonstrating the efficacy and safety of any product candidate in clinical trials or in obtaining regulatory approval thereafter. We also cannot be certain that our scientific and development approach, including with respect to ipsoprubart and ELA822, will result in an approvable or marketable product with commercial value in our initial targeted indications or any other indication.
Further, the pharmaceutical industry is characterized by rapidly advancing technologies, and our future success will depend in part on our ability to maintain a competitive position. If we fail to stay at the forefront of technological change, we may be unable to compete effectively. Our competitors may render our approach obsolete or limit the commercial value of our scientific approach or product candidates through advances in existing technologies or the development of new or different approaches. Conversely, adverse developments with respect to other companies pursuing a similar approach may adversely impact the actual or perceived value of our approach and the potential of our product candidates.
Therefore, we do not know whether our approach, including development of product candidates in immune-mediated diseases and cancer, will be successful, or whether our specific development programs for ipsoprubart and ELA822 will yield approvable or commercially viable products. If our approach or our individual programs are unsuccessful, the value of our company could decline significantly.
Drug development is a lengthy and expensive process, the outcome of clinical testing is inherently uncertain, and results of earlier nonclinical studies and clinical trials may not be predictive of future clinical trial results. We may incur additional costs or experience delays in completing, or ultimately be unable to complete, the development and commercialization of ipsoprubart, ELA822 and any future product candidates for many reasons, including a failure to replicate positive results from earlier nonclinical studies or clinical trials in ongoing or future nonclinical studies or clinical trials.
Before obtaining approval from regulatory authorities for the commercialization of any of our product candidates, we must conduct extensive clinical trials to demonstrate the safety, purity, potency, or efficacy of the product candidate
17
Table of Contents
in humans. Before we can initiate clinical trials for any product candidates, we must submit the results of nonclinical studies to the FDA or comparable foreign regulatory authorities along with other information, including information about product candidate chemistry, manufacturing and controls and our proposed clinical trial protocol, as part of an IND or similar regulatory submission. The FDA or comparable foreign regulatory authorities may require us to conduct additional nonclinical studies for any product candidate before it allows us to initiate clinical trials under any IND or similar regulatory submission, which may lead to delays and increase the costs of our nonclinical development programs. For example, while we have an approved CTA for ELA822 in the EU and initiated a Phase 1 clinical trial, we plan to submit an IND to the FDA for ELA822 in the United States, and the FDA may require additional data before allowing us to proceed.
The risk of failure is high for nonclinical and clinical-stage product candidates. It is impossible to predict when or if ipsoprubart, ELA822 or any future product candidates will receive regulatory approval. To obtain the requisite regulatory approvals to commercialize any product candidate, we must demonstrate through extensive nonclinical studies and lengthy, complex and expensive clinical trials that our product candidates are safe, pure, potent and effective for its claimed indication, which includes clinical effectiveness, in humans. Clinical testing can take many years to complete, and its outcome is inherently uncertain. Failure can occur at any time during the clinical trial process. The results of clinical trials and nonclinical studies of ipsoprubart and ELA822, or any future product candidates may not be predictive of the results of later-stage clinical trials. Interim, topline or preliminary results of a clinical trial are not necessarily indicative of final results. In particular, our ongoing SURPASS Phase 2/3 registrational trial, which is evaluating ipsoprubart in adult and pediatric patients diagnosed with treatment-naïve sHLH, is based primarily on data from 12 frontline mHLH patients enrolled in our Phase 1b trial. The sHLH patient population is heterogeneous, encompassing multiple disease subtypes and underlying triggers, and results observed in a small, single-arm Phase 1b trial may not be reproducible in a larger, more diverse registrational trial, including in pediatric patients, as the Phase 1b trial enrolled adults only. If SURPASS fails to demonstrate substantial evidence of efficacy and sufficient tolerability to satisfy the FDA and other regulators, we may be unable to obtain regulatory approval for ipsoprubart, which would have a material adverse effect on our business, financial condition, results of operations, and prospects.
We may be unable to establish benefit on clinical endpoints that applicable regulatory authorities would consider clinically meaningful, and a clinical trial can fail at any stage of testing. Differences in trial design between early-stage clinical trials and later-stage clinical trials make it difficult to extrapolate the results of earlier clinical trials to later clinical trials. Moreover, clinical data are often susceptible to varying interpretations and analyses, and many companies that have believed their product candidates performed satisfactorily in clinical trials have nonetheless failed to obtain regulatory approval of their products. A number of companies in the biopharmaceutical industry have suffered significant setbacks in advanced clinical trials due to lack of efficacy or to unfavorable safety profiles, notwithstanding promising results in earlier trials. There is typically a high rate of failure of product candidates proceeding through clinical trials, particularly in the earlier stages of development. Most product candidates that commence clinical trials are never approved as products and there can be no assurance that any of our future clinical trials will ultimately be successful or support regulatory approval of ipsoprubart, ELA822 or any future product candidates.
Additionally, our ongoing Phase 2/3 registrational trial of ipsoprubart in sHLH and Phase 1 clinical trial of ipsoprubart in T cell malignancies, and some of our planned clinical trials utilize, or may utilize, an “open-label” trial design. An “open-label” clinical trial is one where both the patient and investigator know whether the patient is receiving the investigational product candidate or either an existing approved drug or placebo. Most typically, open-label clinical trials test only the investigational product candidate and sometimes may do so at different dose levels. Open-label clinical trials are subject to various limitations that may exaggerate any therapeutic effect as patients in open-label clinical trials are aware when they are receiving treatment. Open-label clinical trials may be subject to a “patient bias” where patients perceive their symptoms to have improved merely due to their awareness of receiving an experimental treatment. In addition, open-label clinical trials may be subject to an “investigator bias” where those assessing and reviewing the physiological outcomes of the clinical trials are aware of which patients have received treatment and may interpret the information of the treated group more favorably given this knowledge. The results from an open-label trial may not be predictive of future clinical trial results with any of our product candidates for which we include an open-label clinical trial when studied in a controlled environment with a placebo or active control.
18
Table of Contents
Our SURPASS Phase 2/3 registrational trial for ipsoprubart in sHLH is designed as a single-arm, open-label trial that relies on COMPASS, a retrospective natural history study, to provide the external control comparator rather than a concurrent randomized control group. This design is subject to significant regulatory risk. The FDA or the EMA may disagree with the propensity score weighting methodology used to adjust for differences in baseline characteristics between the SURPASS and COMPASS populations, the adequacy of the COMPASS patient population as a comparator, or the sufficiency of the 8-week OS primary endpoint. Even if SURPASS achieves its primary endpoint in the lymphoma-associated HLH (LA-HLH) population, the FDA or EMA may not accept the data as sufficient to support approval of ipsoprubart for sHLH broadly, and any approval we receive could be limited to a narrower patient population, which could significantly limit the commercial opportunity for ipsoprubart. Retrospective chart review data underlying COMPASS may be of lower quality, completeness, or consistency than prospectively collected data, and the propensity score adjustment may not fully account for all relevant differences in baseline characteristics between the ipsoprubart-treated and external control populations. While we believe we have aligned with the FDA on this approach at the design stage and have engaged with the EMA regarding our development plans, any such alignment or engagement does not guarantee that these agencies will accept the resulting data, even if we believe the results are positive, as sufficient to support regulatory approval. For example, the FDA or EMA may determine that the COMPASS data are insufficient in quality, completeness, or comparability to the SURPASS population, or may require additional data or a randomized controlled trial. If the FDA or EMA does not accept our trial designs or the resulting data as sufficient to support regulatory approval, we may be required to conduct additional or redesigned clinical trials, which would significantly delay and increase the cost of our development program and could prevent approval of ipsoprubart.
We do not know whether any of our nonclinical studies or clinical trials will begin as planned, need to be restructured or be completed on schedule, if at all. We may experience delays in initiating or completing clinical trials. We also may experience numerous unforeseen events during, or as a result of, any future clinical trials that we could conduct that could delay or prevent our ability to receive regulatory approval or commercialize ipsoprubart, ELA822 or any future product candidates, including:
| ∎ | we may be unable to generate sufficient nonclinical, toxicology, or other in vivo or in vitro data to support the initiation or continuation of clinical trials; |
| ∎ | the FDA or comparable foreign regulatory authorities may disagree as to the design or implementation of our clinical trials; |
| ∎ | we may experience delays in identifying, recruiting and training suitable clinical investigators; |
| ∎ | regulatory authorities, institutional review boards (IRBs) or ethics committees may not authorize or allow us or our investigators to commence a clinical trial or conduct a clinical trial at a prospective trial site, or may halt or suspend an ongoing clinical trial; |
| ∎ | we may experience delays in reaching or fail to reach agreement on acceptable terms with prospective trial sites and prospective contract research organizations (CROs) the terms of which can be subject to extensive negotiation and may vary significantly among different CROs and trial sites; |
| ∎ | clinical trial sites deviating from the trial protocol or dropping out of a trial; |
| ∎ | clinical trials of any product candidates may fail to show safety or efficacy, produce negative or inconclusive results and we may decide, or regulatory authorities may require us, to conduct additional nonclinical studies or clinical trials or we may decide to abandon product development programs; |
| ∎ | the number of participants required for clinical trials of any product candidates may be larger than we anticipate, enrollment in these clinical trials may be slower than we anticipate or participants may drop out of these clinical trials or fail to return for post-treatment follow-up at a higher rate than we anticipate; |
| ∎ | our third-party contractors may fail to comply with regulatory requirements or meet their contractual obligations to us in a timely manner, or at all, or may deviate from the clinical trial protocol or drop out of the trial, which may require that we add new clinical trial sites or investigators; |
| ∎ | we may elect, or regulatory authorities, IRBs, or ethics committees may require, that we or our investigators, suspend or terminate clinical research or trials for various reasons, including noncompliance with regulatory requirements or a finding that the patients in our trials are being exposed to unacceptable health risks; |
| ∎ | the cost of clinical trials for ipsoprubart, ELA822 or any future product candidates may be greater than we anticipate, and we may not have sufficient funds to complete such trials; |
19
Table of Contents
| ∎ | the quality of ipsoprubart, ELA822 or any future product candidates or other materials necessary to conduct clinical trials of ipsoprubart, ELA822 or any future product candidates may be inadequate to initiate or complete a given clinical trial; |
| ∎ | our inability to manufacture sufficient quantities of ipsoprubart, ELA822 or any future product candidates for use in clinical trials; |
| ∎ | our inability to meet drug specifications suitable for use in clinical trials and commercial applications, including the development and validation of a potency assay to ensure that the characteristics of the product released are as expected; |
| ∎ | reports from clinical testing of other therapies may raise safety or efficacy concerns about ipsoprubart, ELA822 or any future product candidates; |
| ∎ | the receipt of feedback from regulatory authorities that requires us to modify the design of our clinical trials; |
| ∎ | our failure to establish an appropriate safety profile for a product candidate based on clinical or nonclinical data for such product candidate as well as data emerging from other therapies in the same class as ipsoprubart, ELA822 or any future product candidates; and |
| ∎ | the FDA or other comparable foreign regulatory authorities may require us to submit additional data such as long-term toxicology studies or impose other requirements before permitting us to initiate a clinical trial. |
We could also encounter delays if a clinical trial is suspended or terminated by us, the IRBs or ethics committees overseeing the institutions in which such trials are being conducted, or the FDA or other comparable regulatory authorities, or if a clinical trial is recommended for suspension or termination by the Data Safety Monitoring Board for such trial. A suspension or termination may be imposed due to a number of factors, including failure to conduct the clinical trial in accordance with regulatory requirements, including the FDA’s Good Clinical Practice (GCP) regulations, or our clinical protocols, inspection of the clinical trial operations or trial site by the FDA or other comparable foreign regulatory authorities resulting in the imposition of a clinical hold, unforeseen safety issues or adverse side effects, failure to demonstrate a benefit from using a product or treatment, failure to establish or achieve clinically meaningful trial endpoints, changes in governmental regulations or administrative actions or lack of adequate funding to continue the clinical trial. Clinical trials may also be delayed or terminated as a result of ambiguous or negative interim results. Many of the factors that cause, or lead to, a delay in the commencement or completion of clinical trials may also ultimately lead to the denial of regulatory approval of ipsoprubart, ELA822 or any future product candidates. Further, the FDA or other comparable foreign regulatory authorities may disagree with our clinical trial design and our interpretation of data from clinical trials or may change the requirements for approval even after they have reviewed and commented on the design for our clinical trials.
We cannot predict with any certainty the schedule for commencement and completion of future clinical trials. Further, conducting clinical trials in foreign countries, as we have done and may do in the future for our product candidates, presents additional risks that may delay completion of our clinical trials. These risks include the failure of enrolled patients in foreign countries to adhere to clinical protocol as a result of differences in healthcare services or cultural customs, managing additional administrative burdens associated with foreign regulatory schemes, as well as political and economic risks relevant to such foreign countries.
If we are required to conduct additional clinical trials or other testing of our current or future product candidates beyond those that we currently contemplate, if we are unable to successfully complete clinical trials of our current or future product candidates or other testing in a timely manner, if the results of these trials or tests are not positive or are only modestly positive or if there are safety concerns, we may incur unplanned costs, be delayed in seeking and obtaining regulatory approval, if we receive such approval at all, receive more limited or restrictive regulatory approval, be subject to additional post-marketing testing requirements or have the product removed from the market after obtaining regulatory approval.
Additionally, if the results of our clinical trials are inconclusive or if there are safety concerns or serious adverse events associated with our product candidates, we may:
| ∎ | be delayed in obtaining regulatory approval, if at all; |
| ∎ | obtain approval for indications or patient populations that are not as broad as intended or desired or may have restricted duration expectations or guidance; |
20
Table of Contents
| ∎ | obtain approval with labeling that includes significant use or distribution restrictions or safety warnings; |
| ∎ | be required to perform additional clinical trials to support approval or be subject to additional post-marketing testing requirements; |
| ∎ | have regulatory authorities withdraw, vary or suspend their approval of the product or impose restrictions on its distribution in the form of a Risk Evaluation and Mitigation Strategy (REMS) or comparable foreign strategy; |
| ∎ | be subject to the addition of labeling statements, such as warnings or contraindications; |
| ∎ | be sued; or |
| ∎ | experience damage to our reputation. |
Our drug development costs will also increase if we experience delays in testing or obtaining regulatory approvals. Also, delays in obtaining regulatory approval may increase commercialization costs if the competitive environment becomes more intense prior to market entry.
Moreover, principal investigators for our clinical trials may serve as scientific advisors or consultants to us from time to time and receive compensation in connection with such services. Under certain circumstances, we may be required to report some of these relationships to the FDA or comparable foreign regulatory authorities. The FDA or comparable foreign regulatory authority may conclude that a financial relationship between us and a principal investigator has created a conflict of interest or otherwise affected interpretation of the study. The FDA or comparable foreign regulatory authority may therefore question the integrity of the data generated at the applicable clinical trial site and the utility of the clinical trial itself may be jeopardized. This could result in a delay in approval, or rejection, of our marketing applications by the FDA or comparable foreign regulatory authority, as the case may be, and may ultimately lead to the denial of regulatory approval of one or more of our product candidates.
Furthermore, we may make formulation or manufacturing changes to our product candidates, in which case we may need to conduct additional nonclinical studies to bridge our modified product candidates to earlier versions. If we experience delays in the commencement or completion of our clinical trials, or if we terminate a clinical trial prior to completion, the commercial prospects of our current or any future product candidates could be negatively impacted, and our ability to generate revenues from our current or future product candidates may be delayed or eliminated entirely.
In addition, the FDA’s and other regulatory authorities’ policies with respect to clinical trials may change and additional government regulations may be enacted. For instance, the regulatory landscape related to clinical trials in the EU recently evolved. The EU Clinical Trials Regulation (CTR) which was adopted in April 2014 and repeals the EU Clinical Trials Directive, became applicable on January 31, 2022. While the EU Clinical Trials Directive required a separate CTA to be submitted in each member state in which the clinical trial takes place, to both the competent national health authority and an independent ethics committee, the CTR introduces a centralized process and only requires the submission of a single application for multi-center trials. The CTR allows sponsors to make a single submission to both the competent authority and an ethics committee in each member state, leading to a single decision per member state. The assessment procedure of the CTA has been harmonized as well, including a joint assessment by all member states concerned, and a separate assessment by each member state with respect to specific requirements related to its own territory, including ethics rules. Each member state’s decision is communicated to the sponsor via the centralized EU portal. Once the CTA is approved, clinical study development may proceed. The CTR transition period ended on January 31, 2025, and all clinical trials (and related applications) are now fully subject to the provisions of the CTR. Compliance with the CTR requirements by us and our third-party service providers, such as CROs, may impact our development plans.
If we experience delays or difficulties in the enrollment of patients in clinical trials, our receipt of necessary regulatory approvals could be delayed or prevented.
Patient enrollment is a significant factor in the timing and success of clinical trials, and the timing of our clinical trials depends, in part, on the speed at which we can recruit and enroll patients to participate in our trials, as well as their completion of required follow-up periods.
We may not be able to initiate or continue our ongoing or planned clinical trials for our current or future product candidates if we are unable to identify and enroll a sufficient number of eligible patients to participate in these trials
21
Table of Contents
as required by the FDA or comparable foreign regulatory authorities. We cannot be certain how many patients will meet our criteria for inclusion in our clinical trials, or that the number of patients enrolled in each program will suffice to support an application for regulatory approval in the indications we are targeting. If our strategies for patient identification and enrollment prove unsuccessful, we may have difficulty enrolling or maintaining patients appropriate for our product candidates. Patient enrollment is also affected by other factors, including:
| ∎ | severity of the disease under investigation; |
| ∎ | our ability to recruit clinical trial investigators of appropriate competencies and experience; |
| ∎ | the incidence and prevalence of our target indications; |
| ∎ | clinicians’ and patients’ awareness of, and perceptions as to, the potential advantages and risks of our product candidates in relation to other available therapies, including any new products that may be approved for the indications we are investigating; |
| ∎ | the availability, expertise, dedication and selection of CROs, to manage operations related to clinical trial enrollment; |
| ∎ | competing studies or trials with similar eligibility criteria; |
| ∎ | invasive procedures required to enroll patients and to obtain evidence of the product candidate’s performance during the clinical trial; |
| ∎ | availability and efficacy of approved medications for the disease under investigation; |
| ∎ | eligibility criteria defined in the protocol for the trial in question; |
| ∎ | the size and nature of the patient population required for analysis of the trial’s primary endpoints; |
| ∎ | efforts to facilitate timely enrollment in clinical trials; |
| ∎ | whether we are subject to a partial or full clinical hold on any of our clinical trials; |
| ∎ | reluctance of physicians or patient advocacy organizations to encourage patient participation in clinical trials; |
| ∎ | the ability to monitor patients adequately during and after treatment; |
| ∎ | our ability to obtain and maintain patient consents; and |
| ∎ | proximity and availability of clinical trial sites for prospective patients. |
Furthermore, we expect to rely on our CROs and clinical trial sites to ensure the proper and timely conduct of our future clinical trials, including the patient enrollment process, and we have limited influence over their performance. Additionally, we could encounter delays if treating physicians face unresolved ethical issues associated with enrolling patients in future clinical trials of ipsoprubart, ELA822 or any future product candidates in lieu of prescribing existing treatments that have established safety and efficacy profiles.
If we are unable to enroll a sufficient number of patients for our clinical trials, it would result in significant delays or might require us to abandon one or more clinical trials altogether. Even if we are able to enroll a sufficient number of patients for our clinical trials, we may have difficulty maintaining patients in our clinical trials. Enrollment delays in our clinical trials may result in increased development costs for ipsoprubart, ELA822 or any future product candidates, slow down or halt our product candidate development and approval process and jeopardize our ability to seek and obtain the regulatory approval required to commence product sales and to generate revenue, which would cause our stock price to decline and limit our ability to obtain additional financing, if needed.
Adverse side effects or other safety risks associated with ipsoprubart, ELA822 or any future product candidates we may develop could delay or preclude approval, cause us to suspend or discontinue clinical trials or abandon further development, limit the commercial profile of an approved product, or result in significant negative consequences following regulatory approval, if any.
There may be treatment-related SAEs or unexpected serious adverse reactions suspected to be associated with the use of ipsoprubart, ELA822 or any future product candidates. Our clinical trials may reveal significant adverse events not seen in our nonclinical studies or prior clinical trials and may result in a safety or tolerability profile that could delay or prevent regulatory approval, or market acceptance of ipsoprubart, ELA822 or any future product candidates, if approved. Undesirable or clinically unmanageable side effects observed in our clinical trials for our product candidates could occur and cause us or regulatory authorities to interrupt, delay or halt our clinical trials
22
Table of Contents
and could result in more restrictive labeling than anticipated or the delay or denial of regulatory approval by the FDA or comparable foreign regulatory authorities. If additional adverse events, SAEs or other side effects are observed in any of our clinical trials that are atypical of, or more severe than, the known side effects of the respective class of agents that each of our product candidates are a part of, we may have difficulty recruiting patients to our clinical trials, patients may drop out of our trials, or we may be required to abandon those trials or our development efforts of one or more product candidates altogether. Moreover, we, the FDA, other comparable regulatory authorities or an IRB may suspend clinical trials of a product candidate at any time for various reasons, including a belief that participants in such trials are being exposed to unacceptable health risks or adverse side effects. Even if the side effects do not preclude the product candidate from obtaining or maintaining regulatory approval, undesirable side effects may inhibit market acceptance due to tolerability concerns as compared to other available therapies. Any of these developments could materially harm our business, financial condition and prospects.
Furthermore, clinical trials by their nature utilize a sample of the potential patient population. With a limited number of participants and limited duration of exposure, rare and severe side effects of our product candidates or those of our competitors may only be uncovered with a significantly larger number of patients exposed to the drug. Undesirable or clinically unmanageable side effects observed in our clinical trials for our product candidates could also occur following discontinuation of ipsoprubart, ELA822, or any future product candidates with sufficient recovery periods, and we will need to monitor the severity and duration of side effects in our clinical trials. If such effects are more severe, less reversible than we expect or not reversible at all, we may decide or be required to perform additional studies or to halt or delay further clinical development of ipsoprubart and ELA822, as applicable, or any clinical or preclinical development of future product candidates, which could result in the delay or denial of regulatory approval by the FDA or other comparable foreign regulatory authorities. Adverse events and SAEs that emerge during clinical investigation of or treatment with ipsoprubart, ELA822 or any future product candidates may be deemed to be related to our product candidates.
Additionally, if any of our product candidates receives regulatory approval, and we or others later identify undesirable side effects caused by such product, a number of potentially significant negative consequences could result. For example, the FDA could require us to adopt a REMS, which may include, among other things, a communication plan to health care practitioners, patient education, extensive patient monitoring or distribution systems and processes that are highly controlled, restrictive and more costly than what is typical for the industry. We may also be required to adopt a REMS or engage in similar actions, such as patient education, certification of health care professionals or specific monitoring, if we or others later identify undesirable side effects caused by any product that we develop alone. Other potentially significant negative consequences associated with adverse events include:
| ∎ | we may be required to suspend marketing of a product, or we may decide to remove such product from the marketplace; |
| ∎ | regulatory authorities may withdraw or change their approvals of a product; |
| ∎ | regulatory authorities may require additional warnings on the label or limit access of a product to selective specialized centers with additional safety reporting and with requirements that patients be geographically close to these centers for all or part of their treatment; |
| ∎ | we may be required to create a medication guide outlining the risks of a product for patients, or to conduct post-marketing studies; |
| ∎ | we may be required to change the way a product is administered; |
| ∎ | we could be subject to fines, injunctions, or the imposition of criminal or civil penalties, or be sued and held liable for harm caused to participants or patients; and |
| ∎ | a product may become less competitive, and our reputation may suffer. |
Any of these events could diminish the usage or otherwise limit the commercial success of our product candidates and prevent us from achieving or maintaining market acceptance of our product candidates, if approved by the FDA or other regulatory authorities.
23
Table of Contents
Preliminary, topline or interim data from our clinical trials that we announce or publish from time to time may change as more patient data become available and/or are subject to audit and verification procedures that could result in material changes in the final data.
From time to time, we may publicly disclose preliminary, topline or interim data from our clinical trials, such as preliminary, topline or interim data analysis from our ongoing Phase 2/3 registrational trial of ipsoprubart in sHLH and Phase 1 clinical trial of ipsoprubart in T cell malignancies. These data and related findings and conclusions may only reflect certain endpoints rather than all endpoints and are subject to change. We also make assumptions, estimations, calculations and conclusions as part of our analyses of data, and we may not have received or had the opportunity to fully and carefully evaluate all data. As a result, the preliminary or topline results that we report may differ from future results of the same studies, or different conclusions or considerations may qualify such results, once additional data have been received and fully evaluated.
Preliminary or topline data also remain subject to review and verification procedures that may result in the final data being materially different from the preliminary or topline data we previously published. As a result, preliminary and topline data should be viewed with caution until the final data are available. In addition, we may report preliminary data or interim analyses of the clinical trials we may conduct and complete, which are subject to the risk that one or more of the clinical outcomes may materially change as patient enrollment continues and more patient data become available. Adverse changes between preliminary or interim data and final data could significantly harm our business and prospects. Further, additional disclosure of preliminary or interim data by us in the future could result in volatility in the price of our common stock.
Further, the information we choose to publicly disclose regarding a particular study or clinical trial is typically selected from a more extensive amount of available information. You or others may not agree with what we determine is the material or otherwise appropriate information to include in our disclosure, and any information we determine not to disclose may ultimately be deemed significant with respect to future decisions, conclusions, views, activities or otherwise regarding a particular product, product candidate or our business. If the preliminary, topline or interim data that we report differ from later, final or actual results, or if others, including regulatory authorities, disagree with the conclusions reached, our ability to obtain approval for, and commercialize, our product candidates may be harmed, which could harm our business, financial condition, results of operations, and prospects.
The incidence and prevalence for target patient populations of one or more of our current product candidates have not been established with precision. If the market opportunities for our current or future product candidates are smaller than we estimate or if any approval that we may obtain is based on a narrower definition of the patient population, our revenue and ability to achieve profitability will be adversely affected, possibly materially.
Our projections of the number of people who have sHLH and T cell malignancies, as well as other diseases in immunology and cancer that we are targeting, and who have the potential to benefit from treatment with ipsoprubart, ELA822 or any of our future product candidates, are based on our beliefs and estimates. These estimates have been derived from a variety of sources, including scientific literature, surveys of clinics, patient foundations or market research, and may prove to be incorrect. In particular, our estimates of the sHLH patient population and projected growth rates are based primarily on a single internally commissioned claims data analysis using International Classification of Diseases diagnosis codes, which may not capture misdiagnosed or undiagnosed patients and may be subject to coding variability. The compound annual growth rate we have cited for the sHLH diagnosed population may not be sustained, and the projected patient population may be materially lower than our estimates. Further, new studies may change the estimated incidence or prevalence of the indications that we are targeting. The potentially addressable patient population for ipsoprubart, ELA822 or any of our future product candidates may be more limited than we currently estimate or may not be amenable to treatment with such product candidates.
Although we intend to explore other therapeutic opportunities in addition to the product candidates that we are currently developing, we may fail to identify viable new product candidates for clinical development for a number of reasons. If we fail to identify additional potential product candidates, our business could be materially harmed.
We may expend our limited resources to pursue a particular product candidate in specific target indications and fail to capitalize on product candidates or indications that may be more profitable or for which we believe there is a greater likelihood of success. Because we have limited financial and managerial resources, we focus our development efforts on certain selected product candidates in certain selected target indications. For example, we
24
Table of Contents
are currently primarily focused on ipsoprubart for the treatment of sHLH and T cell malignancies. As a result, we may forgo or delay pursuit of opportunities with other product candidates, or other indications for our current or any future product candidates that later prove to have greater commercial potential. Our resource allocation decisions may cause us to fail to capitalize on viable commercial products or profitable market opportunities. Our spending on current and future development programs and product candidates for specific target indications may not yield any commercially viable product candidates. If we do not accurately evaluate the commercial potential or target market for a particular product candidate, we may relinquish valuable rights to that product candidate through collaboration, licensing or other royalty arrangements in cases in which it would have been more advantageous for us to retain sole development and commercialization rights to such product candidate.
We are conducting and may in the future conduct clinical trials for current or future product candidates outside the United States, and the FDA and comparable foreign regulatory authorities may not accept data from such trials.
We are conducting and may in the future conduct, clinical trials for current or future product candidates outside the United States, and the FDA and comparable foreign regulatory authorities may not accept data from such trials. We are currently conducting clinical trials in the EU, and we expect to continue to conduct trials internationally in the future. The acceptance of data from clinical trials conducted outside the United States by the FDA or other comparable foreign regulatory authorities may be subject to certain conditions or may not be accepted at all. In cases where data from foreign clinical trials are intended to serve as the sole basis for regulatory approval in the United States, the FDA will generally not approve the application unless (i) the data are applicable to the U.S. population and U.S. medical practice and (ii) the trials were performed by clinical investigators of recognized competence and pursuant to GCP regulations, and the data may be considered valid without the need for an on-site inspection by the FDA or, if the FDA considers such an inspection to be necessary, the FDA is able to validate the data through an on-site inspection or other appropriate means. In addition, even where the foreign study data are not intended to serve as the sole basis for approval, if the study was not otherwise conducted pursuant to an IND, the FDA will not accept the data as support for an IND or marketing application unless the study is well-designed and well-conducted in accordance with GCPs and the FDA is able to validate the data from the study through an onsite inspection if deemed necessary. Additionally, the FDA’s clinical trial requirements, including sufficient size of patient populations and statistical powering, must be met. Many foreign regulatory authorities have similar approval requirements. In addition, such foreign trials are subject to the applicable local laws of the foreign jurisdictions where the trials are conducted. Conducting clinical trials outside the United States also exposes us to additional risks, including risks associated with foreign exchange fluctuations, compliance with foreign manufacturing, customs, shipment and storage requirements, and cultural differences in medical practice and clinical research, and diminished protection of intellectual property in some countries.
If the FDA or other comparable foreign regulatory authorities do not accept such data, it would result in the need for additional trials, which could be costly and time-consuming, and which may result in current or future product candidates that we may develop being delayed or not receiving approval for commercialization in the applicable jurisdiction.
If, in the future, we are unable to establish sales and marketing capabilities or enter into agreements with third parties to sell and market any product we may develop, we may not be successful in commercializing those products if they are approved.
We do not have a sales or marketing infrastructure and have limited experience in the sales, marketing and distribution of drug products. To achieve commercial success for any approved product, we must either develop a sales and marketing organization or outsource these functions to third parties. In the future and if any of our product candidates are approved, we may choose to build a focused sales, marketing and commercial support infrastructure to sell, or participate in sales activities with collaborators for some of our current or future product candidates.
There are risks involved with both establishing our own commercial capabilities and entering into arrangements with third parties to perform these services. For example, factors that may inhibit our efforts to commercialize any approved product candidates include:
| ∎ | the inability to recruit and retain adequate numbers of effective sales, marketing, coverage or reimbursement, customer service, medical affairs and other support personnel; the inability of sales personnel to obtain access to or educate decision makers of the utility of future approved product candidates; |
25
Table of Contents
| ∎ | the inability of reimbursement professionals to negotiate arrangements for formulary access, reimbursement and other acceptance by payors; |
| ∎ | the inability to price any of our current or future product candidates at a sufficient price point to ensure an adequate and attractive level of profitability; |
| ∎ | restricted or closed distribution channels that make it difficult to distribute our current or future product candidates to segments of the patient population; |
| ∎ | the lack of complementary product candidates to be offered by sales personnel, which may put us at a competitive disadvantage relative to companies with more extensive product candidate lines; and |
| ∎ | unforeseen costs and expenses associated with creating an independent commercialization organization. |
If the commercial launch of a product candidate, if approved, for which we recruit a sales force and establish marketing and other commercialization capabilities is delayed or does not occur for any reason, we would have prematurely or unnecessarily incurred these commercialization expenses. This may be costly, and our investment would be lost if we cannot retain or reposition our commercialization personnel.
If we enter into arrangements with third parties to perform sales, marketing, commercial support and distribution services, our sales revenue or the profitability of sales revenue may be lower than if we were to do so ourselves. In addition, we may not be successful in entering into arrangements with third parties to commercialize our product candidates or may be unable to do so on terms that are favorable to us. We may have little control over such third parties, and any of them may fail to devote the necessary resources and attention to sell and market our product candidates effectively. If we do not establish commercialization capabilities successfully, either on our own or in collaboration with third parties, we will not be successful in commercializing our product candidates, if approved.
Our current or future product candidates may not achieve adequate market acceptance among physicians, patients or their families, healthcare payors and others in the medical community necessary for commercial success.
Even if our current or future product candidates receive regulatory approval, they may not gain adequate market acceptance among physicians, patients or their families, third-party payors and others in the medical community. The degree of market acceptance of any of our approved product candidates will be dependent on a number of factors, including:
| ∎ | the efficacy, durability and safety profile as demonstrated in clinical trials compared to alternative treatments; |
| ∎ | the timing of market introduction of the product candidate as well as competitive products; |
| ∎ | the clinical indications for which a product candidate is approved; |
| ∎ | restrictions on the use of product candidates in the labeling approved by regulatory authorities, such as boxed warnings or contraindications in labeling, or a REMS or comparable foreign strategy, if any, which may not be required of alternative treatments and competitor products; |
| ∎ | the terms of any approvals and the countries in which such approvals are obtained; |
| ∎ | the potential and perceived advantages of our current or future product candidates over alternative treatments; |
| ∎ | the cost of treatment in relation to alternative treatments and the cost/benefit ratios of each; |
| ∎ | the availability of coverage and adequate reimbursement by third-party payors, including government authorities, and timing of relevant formulary decision-making resulting in this coverage and reimbursement; |
| ∎ | the availability of an approved product candidate for use as a combination therapy; |
| ∎ | relative convenience and ease of administration in relation to competition; |
| ∎ | the willingness of the target patient population to try new therapies and of physicians to prescribe these therapies; |
| ∎ | the effectiveness of sales, marketing efforts and market access; |
| ∎ | publicity relating to our product candidates or those of our competitors; |
| ∎ | potential product liability claims; and |
| ∎ | the approval of new therapies for the same indications. |
26
Table of Contents
If any of our current or future product candidates are approved but do not achieve an adequate level of acceptance by physicians, hospitals, healthcare payors and patients, we may not generate or derive sufficient revenue from that product candidate and our financial results could be negatively impacted. Our efforts to educate the medical community and third-party payors regarding the benefits of our products may require significant resources and may never be successful.
If our product candidates do not achieve projected development milestones or commercialization in the announced or expected timeframes, the further development or commercialization of such product candidates may be delayed, and our business will be harmed.
We have estimated, and may in the future estimate, the timing of the accomplishment of various scientific, clinical, manufacturing, regulatory and other product development objectives. These milestones have and may include our expectations regarding the commencement or completion of nonclinical studies and clinical trials, data readouts, the submission of regulatory filings, the receipt of regulatory approval or the realization of other commercialization objectives. The achievement of many of these milestones may be outside of our control. All of these milestones are based on a variety of assumptions, including assumptions regarding capital resources, constraints and priorities, progress of and results from development activities and the receipt of key regulatory approvals or actions, any of which may cause the timing of achievement of the milestones to vary considerably from our estimates. If we or our collaborators fail to achieve announced milestones in the expected timeframes, the commercialization of the product candidates may be delayed, our credibility may be undermined, our business and results of operations may be harmed and the trading price of our common stock may decline.
Risks Related to Our Business and Operations
Our future performance is dependent on our ability to retain key employees and to attract, retain and motivate qualified personnel and manage our human capital.
Our ability to compete in the highly competitive biotechnology and biopharmaceutical industries is largely dependent on our ability to attract, motivate and retain highly qualified managerial, clinical, quality control, scientific and medical personnel. We are highly dependent on the development and management expertise of our executive officer team. We currently do not maintain “key person” life insurance on these individuals or any of our employees. This lack of insurance means that we may not have adequate compensation for the loss of the services of these individuals. The loss of one or more members of our management team or other key employees or advisors could delay our research and development programs and have a material and adverse effect on our business, financial condition, results of operations, and prospects. We are dependent on the continued service of our technical personnel, because of the highly technical nature of ipsoprubart, ELA822 or any future product candidates and technologies, and the specialized nature of the regulatory approval process. Because our management team and key employees are not obligated to provide us with continued service, they could terminate their employment with us at any time without penalty.
In addition, job candidates and existing employees often consider the value of the stock awards they receive in connection with their employment. If the perceived benefits of our stock awards decline, either because we are a public company or for other reasons, it may harm our ability to recruit and retain highly skilled employees. Our employees may be more likely to leave us if the shares they own have significantly appreciated in value relative to the original purchase prices of the shares, or if the exercise prices of the options that they hold are significantly below the market price of our common stock, particularly after the expiration of the lock-up agreements described herein.
We conduct our in-person operations at our facilities in South San Francisco, California. This region is a headquarter to many other biopharmaceutical companies and academic and research institutions. Competition for skilled personnel in our market, and nationally, is intense and may limit our ability to hire and retain highly qualified personnel on acceptable terms or at all. We also face competition for personnel from other companies, universities, public and private research institutions, government entities and other organizations. Our industry has experienced a high rate of turnover of management personnel in recent years. Our future performance will be dependent in large part on our continued ability to attract and retain highly qualified scientific, technical and management personnel, as well as personnel with expertise in clinical testing, manufacturing, governmental regulation and
27
Table of Contents
commercialization. If we are unable to continue to attract and retain high-quality personnel, the rate and success at which we can discover and develop product candidates will be limited, which could have a material and adverse effect on our business, financial condition, results of operations, and prospects.
Our future growth may be dependent, in part, on our ability to operate in foreign markets, where we would be subject to additional regulatory burdens and other risks and uncertainties.
Our future growth may be dependent, in part, on our ability to develop and commercialize ipsoprubart and ELA822, if approved, and any future product candidates in foreign markets for which we may rely on collaboration with third parties. We are not permitted to market or promote ipsoprubart, ELA822 or any future product candidates before we receive regulatory approval from the applicable regulatory authority in that foreign market and may never receive such regulatory approval for ipsoprubart, ELA822 or any future product candidates. To obtain separate regulatory approval in many other countries, we must comply with numerous and varying regulatory requirements of such countries regarding safety and efficacy and governing, among other things, clinical trials and commercial sales, pricing and distribution of ipsoprubart, ELA822 or any future product candidates, and we cannot predict success in these jurisdictions. Approval procedures may be more onerous than those in the United States and may require that we conduct additional nonclinical studies or clinical trials. If we fail to comply with the regulatory requirements in international markets and receive applicable regulatory approvals, our target market will be reduced and our ability to realize the full market potential of ipsoprubart, ELA822 or any future product candidates will be harmed and our business will be adversely affected. We may not obtain foreign regulatory approvals on a timely basis, if at all. Our failure to obtain approval of any of ipsoprubart, ELA822 or any future product candidates by regulatory authorities in another country may significantly diminish the commercial prospects of that product candidate and our business, financial condition, results of operations, and prospects could be materially and adversely affected. Moreover, even if we obtain approval of ipsoprubart, ELA822 or any future product candidates and ultimately commercialize ipsoprubart, ELA822 or any future product candidates in foreign markets, we would be subject to additional risks and uncertainties, including the burden of complying with complex and changing foreign regulatory, tax, accounting and legal requirements and reduced protection of intellectual property rights in some foreign countries.
We expect to expand our development, clinical and regulatory capabilities and operations as we grow, and as a result, we may encounter difficulties in managing our growth, which could disrupt our operations.
As of June 30, 2026, we had 46 full-time employees. We expect to increase the number of our employees and the scope of our operations, particularly in the areas of clinical development, clinical operations, manufacturing, late-stage regulatory affairs, finance, accounting, management information systems, business operations, public company compliance, communications and other corporate development functions, and, if ipsoprubart, ELA822 or any of our future product candidates receive regulatory approval, sales, marketing and distribution capabilities. If we acquire additional product candidates or enter into collaborations in the future, we may have to further expand our employee base beyond our current projections, which may include further preclinical research and development or additional later-stage regulatory operations. To manage our anticipated future growth, we must continue to implement and improve our managerial, operational and financial systems, expand our facilities and continue to recruit and train additional qualified personnel. Due to our limited financial resources and the complexities associated with managing anticipated growth and developing sales, marketing, and distribution infrastructure, our management team may encounter challenges in scaling our operations efficiently, including recruiting, training, and retaining additional qualified personnel. As our operations expand, we also expect that we will need to manage additional relationships with various strategic partners, suppliers and other third parties. The expansion of our operations may lead to significant costs and may divert the time and attention of our management from day-to-day activities and harm our development efforts. If we are unable to raise capital when needed or on acceptable terms, we may be forced to delay, reduce, or eliminate certain of our research and development programs.
If we are not able to effectively manage growth and expand our operations, we may not be able to successfully implement the tasks necessary to further develop and commercialize, if approved, ipsoprubart, ELA822 or any future product candidates and, accordingly, we may not achieve our research, development and commercialization goals.
28
Table of Contents
We face substantial competition, which may result in others discovering, developing or commercializing products before or more successfully than we do.
The development and commercialization of new drug products is highly competitive. We face competition from entities that have made substantial investments into the rapid development of novel treatments for disorders associated with immune-mediated diseases and cancer, including large and specialty biopharmaceutical and biotechnology companies, some of which already have approved therapies in our current indications. The development and commercialization of product candidates for disorders associated with immune-mediated diseases and cancer is highly competitive. Our current and any future product candidates, if approved, will face significant competition, including from well-established, currently marketed therapies or recommended standards of care, and our failure to demonstrate a meaningful improvement to the existing standards of care may prevent us from achieving significant market penetration. Many of our competitors have significantly greater resources and experience than we do and we may not be able to successfully compete. We face substantial competition from multiple sources, including large and specialty biopharmaceutical and biotechnology companies, academic research institutions and governmental agencies and public and private research institutions. In particular, for ipsoprubart in sHLH, although there are no therapies approved for the treatment of sHLH broadly, clinicians currently rely on a heterogeneous combination of off-label agents, including corticosteroids, etoposide, alemtuzumab, ruxolitinib, tocilizumab, anakinra, and, in certain subtypes, emapalumab, which is approved for primary HLH and macrophage activation syndrome associated with Still’s disease but not for sHLH broadly. These agents are familiar to treating physicians, are generally available at low cost, and are supported by institutional protocols and clinical experience accumulated over many years. Even if ipsoprubart receives regulatory approval, physicians may prefer to continue using these familiar off-label regimens rather than adopting a newly approved therapy, particularly given the acute and rapidly evolving nature of sHLH, which may limit the time available for physicians to evaluate new treatment options. If we are unable to demonstrate to physicians, patients, and payors that ipsoprubart offers a meaningful clinical advantage over existing off-label approaches, or if physicians are reluctant to change established treatment practices, our ability to achieve commercial success for ipsoprubart would be materially impaired.
Our current product candidates, initially under development for treatment of various immune-mediated diseases and cancer, if approved, would face competition from approved treatments, some of which have achieved commercial success. To compete successfully, we need to differentiate our product candidates from these currently marketed drugs, meaning that we will have to demonstrate that the relative cost, method of administration, safety, tolerability or efficacy of our product candidates provides a better alternative to existing and new therapies. Our commercial opportunity and likelihood of success will be reduced or eliminated if our product candidates are not ultimately demonstrated to be safer, more effective, more conveniently administered or less expensive than the current standards of care. Furthermore, even if our product candidates are able to achieve these attributes, acceptance of our products may be inhibited by the reluctance of physicians to switch from existing therapies to our products, or if physicians choose to reserve our products for use in limited circumstances.
Many of our competitors have significantly greater financial, technical, manufacturing, marketing, sales and supply resources or experience than us. If we obtain regulatory approval for any product candidate, we will face competition based on many different factors, including the safety and effectiveness of our current or any future product candidates, the ease with which our current or any future product candidates can be administered and the extent to which patients accept relatively new routes of administration, the timing and scope of regulatory approvals for these product candidates, the availability and cost of manufacturing, marketing and sales capabilities, price, reimbursement coverage and patent position. Competing products could present superior treatment alternatives, including by being more effective, safer, less expensive or marketed and sold more effectively than any products we may develop. Competitive products may make any products we develop obsolete or noncompetitive before we recover the expense of developing and commercializing our current or any future product candidates. Such competitors could also recruit our employees, which could negatively impact our level of expertise and our ability to execute our business plan. In addition, any collaborators may decide to market and sell products that compete with the product candidates that we have agreed to license to them, and any competition by our collaborators could also have a material adverse effect on our future business, financial condition, results of operations, and prospects. Mergers and acquisitions in the biopharmaceutical and biotechnology industries may result in even more resources being concentrated among a smaller number of our competitors. Smaller or earlier-stage companies may also prove to be significant competitors, particularly through collaborative arrangements with large and established companies. These
29
Table of Contents
third parties compete with us in recruiting and retaining qualified management and other personnel and establishing clinical trial sites and recruiting participants for clinical trials, as well as in acquiring technologies complementary to, or necessary for, our programs.
If our information technology systems or those third parties with whom we work or our data, are or were compromised, we could experience adverse consequences resulting from such compromise, including but not limited to regulatory investigations or actions; litigation; fines and penalties; disruptions of our business operations; reputational harm; loss of revenue or profits; and other adverse consequences.
In the ordinary course of our business, we and the third parties with whom we work, collect, receive, store, process, generate, use, transfer, disclose, make accessible, protect, secure, dispose of, transmit, and share (collectively, process) proprietary, confidential, and sensitive data, including personal data (such as health-related data), intellectual property, trade secrets and clinical trial data (collectively, sensitive information).
Cyber-attacks, malicious internet-based activity, online and offline fraud, and other similar activities threaten the confidentiality, integrity, and availability of our sensitive information and information technology systems, and those of the third parties with whom we work. Such threats are prevalent and continue to rise, are increasingly difficult to detect, and come from a variety of sources, including traditional computer “hackers,” threat actors, “hacktivists,” organized criminal threat actors, personnel (such as through theft or misuse), sophisticated nation states, and nation-state-supported actors.
Some actors now engage and are expected to continue to engage in cyber-attacks, including without limitation nation-state actors for geopolitical reasons and in conjunction with military conflicts and defense activities. During times of war and other major conflicts, we and the third parties with whom we work may be vulnerable to a heightened risk of these attacks, including retaliatory cyber-attacks, that could materially disrupt our systems and operations, supply chain, and ability to produce, sell and distribute our goods and services.
We and the third parties with whom we work are subject to a variety of evolving threats, including but not limited to social-engineering attacks (including through deep fakes, which may be increasingly more difficult to identify as fake, and phishing attacks), malicious code (such as viruses and worms), malware (including as a result of advanced persistent threat intrusions), denial-of-service attacks, credential stuffing attacks, credential harvesting, personnel misconduct or error, ransomware attacks, supply-chain attacks, software bugs, server malfunctions, software or hardware failures, loss of data or other information technology assets, adware, telecommunications failures, earthquakes, fires, floods, attacks enhanced or facilitated by AI, and other similar threats.
In particular, severe ransomware attacks are becoming increasingly prevalent and can lead to significant interruptions in our operations, ability to provide our products or services, loss of sensitive data and income, reputational harm, and diversion of funds. Extortion payments may alleviate the negative impact of a ransomware attack, but we may be unwilling or unable to make such payments due to, for example, applicable laws or regulations prohibiting such payments.
It may be difficult and/or costly to detect, investigate, mitigate, contain, and remediate a security incident. Our efforts to do so may not be successful. Actions taken by us or the third parties with whom we work to detect, investigate, mitigate, contain, and remediate a security incident could result in outages, data losses, and disruptions of our business. Threat actors may also gain access to other networks and systems after a compromise of our networks and systems. For example, threat actors may use an initial compromise of one part of our environment to gain access to other parts of our environment or leverage a compromise of our networks or systems to gain access to the networks or systems of third parties with whom we work, such as through phishing or supply chain attacks. There can also be no assurance that our and our third party service providers’, strategic partners’, contractors’, consultants’, CROs’ and collaborators’ cybersecurity risk management program and processes, including our policies, controls or procedures, will be fully implemented, complied with or effective in protecting our systems and sensitive information.
Remote work has increased risks to our information technology systems and data, as our employees utilize network connections, computers and devices outside our premises or network, including working at home, while in transit and in public locations. Additionally, any integration of artificial intelligence in our or any third party’s operations, products or services is expected to pose new or unknown cybersecurity risks and challenges.
30
Table of Contents
Future or past business transactions (such as acquisitions or integrations) could expose us to additional cybersecurity risks and vulnerabilities, as our systems could be negatively affected by vulnerabilities present in acquired or integrated entities’ systems and technologies. Furthermore, we may discover security issues that were not found during due diligence of such acquired or integrated entities, and it may be difficult to integrate companies into our information technology environment and security program.
We rely on third parties to operate critical business systems to process sensitive information in a variety of contexts, including, without limitation, cloud-based infrastructure, data center facilities, employee email, and other functions. Our ability to monitor these third parties’ information security practices is limited, and these third parties may not have adequate information security measures in place. If the third parties with whom we work experience a security incident or other interruption, we could experience adverse consequences. While we may be entitled to damages if the third parties with whom we work fail to satisfy their privacy or security-related obligations to us, any award may be insufficient to cover our damages, or we may be unable to recover such award. In addition, supply-chain attacks have increased in frequency and severity, and we cannot guarantee that third parties’ infrastructure in our supply chain or that of the third parties with whom we work have not been compromised.
While we have implemented security measures designed to protect against security incidents, there can be no assurance that these measures will be effective. We take steps designed to detect, mitigate, and remediate vulnerabilities in our information systems (such as our hardware and/or software, including that of third parties with whom we work). For example, in May 2026, an unknown third party exploited a vulnerability in a third-party plugin for our website hosting software and redirected our website visitors to other websites. While we were able to detect this incident and respond promptly, we may not detect and remediate all such vulnerabilities on a timely basis. Further, we may experience delays in developing and deploying remedial measures and patches designed to address identified vulnerabilities. Vulnerabilities could be exploited and result in a security incident.
Any of the previously identified or similar threats may cause a security incident or other interruption that may result in unauthorized, unlawful, or accidental acquisition, modification, destruction, loss, alteration, encryption, disclosure of, or access to our sensitive information or our information technology systems, or those of the third parties with whom we work. A security incident or other interruption could disrupt our ability (and that of third parties with whom we work) to provide our platform/products/services. We may expend significant resources or modify our business activities (including our clinical trial activities) to try to protect against security incidents. Certain data privacy and security obligations have required us to implement and maintain specific security measures or industry-standard or reasonable security measures to protect our information technology systems and sensitive information.
Applicable data privacy and security obligations may require us, or we may voluntarily choose, to notify relevant stakeholders, including affected individuals, CROs, regulators, and investors, of security incidents, or to take other actions, such as providing credit monitoring and identity theft protection services. Such disclosures and related actions can be costly, and the disclosure or the failure to comply with such applicable requirements could lead to adverse consequences.
In addition to experiencing a security incident, third parties may gather, collect, or infer sensitive information about us from public sources, data brokers, or other means that reveals competitively sensitive details about our organization and could be used to undermine our competitive advantage or market position. Additionally, sensitive information of the Company could be leaked, disclosed, or revealed as a result of or in connection with our employees’, personnel’s, or vendors’ use of generative AI technologies.
If we (or a third party with whom we work) experience a security incident or are perceived to have experienced a security incident, we may experience material adverse consequences, such as government enforcement actions (for example, investigations, fines, penalties, audits, and inspections); additional reporting requirements and/or oversight; restrictions on processing sensitive information (including personal data); incident response, system restoration or remediation and future compliance costs; litigation (including class action claims); indemnification obligations; negative publicity; reputational harm; monetary fund diversions; diversion of management attention; interruptions in our operations (including availability of data); financial loss; and other similar harms. Security incidents and attendant material consequences may negatively impact our ability to grow and operate our business.
31
Table of Contents
Our contracts may not contain limitations of liability, and even where they do, there can be no assurance that limitations of liability in our contracts are sufficient to protect us from liabilities, damages, or claims related to our data privacy and security obligations. We cannot be sure that our insurance coverage will be adequate or sufficient to protect us from or to mitigate liabilities arising out of our privacy and security practices, that such coverage will continue to be available on commercially reasonable terms or at all, or that such coverage will pay future claims.
We and the third parties with whom we work are subject to stringent and evolving U.S. and foreign laws, regulations, and rules, contractual obligations, industry standards, policies and other obligations related to data privacy and security. Our (or the third parties with whom we work) actual or perceived failure to comply with such obligations could lead to regulatory investigations or actions; litigation (including class claims) and mass arbitration demands; fines and penalties; disruptions of our business operations; reputational harm; loss of revenue or profits; and other adverse business consequences.
In the ordinary course of business, we process personal data and other sensitive information. Our data processing activities subject us to numerous data privacy and security obligations, such as various laws, regulations, guidance, industry standards, external and internal privacy and security policies, contractual requirements, and other obligations relating to data privacy and security.
In the United States, federal, state, and local governments have enacted numerous data privacy and security laws, including data breach notification laws, personal data privacy laws, consumer protection laws (e.g., Section 5 of the Federal Trade Commission Act), and other similar laws (e.g., wiretapping laws). For example, the federal Health Insurance Portability and Accountability Act of 1996 (HIPAA), as amended by the Health Information Technology for Economic and Clinical Health Act (HITECH), imposes specific requirements relating to the privacy, security, and transmission of individually identifiable protected health information. We may obtain health information from third parties, such as research institutions with which we collaborate, that are subject to privacy and security requirements under HIPAA. Although we do not believe that we are directly subject to HIPAA, other than potentially with respect to providing certain employee benefits, we could be subject to criminal penalties if we knowingly obtain or disclose individually identifiable health information maintained by a HIPAA covered entity in a manner that is not authorized or permitted by HIPAA.
Numerous U.S. states have enacted comprehensive privacy laws that impose certain obligations on covered businesses, including providing specific disclosures in privacy notices and affording residents with certain rights concerning their personal data. As applicable, such rights may include the right to access, correct, or delete certain personal data, and to opt-out of certain data processing activities, such as targeted advertising, profiling, and automated decision-making. The exercise of these rights may impact our business and ability to provide our products and services. Certain states also impose stricter requirements for processing certain personal data, including sensitive information, such as conducting data privacy impact assessments. These state laws allow for statutory fines for noncompliance. For example, the California Consumer Privacy Act of 2018 (CCPA) applies to personal data of consumers, business representatives, and employees who are California residents, and requires businesses to provide specific disclosures in privacy notices and honor requests of such individuals to exercise certain privacy rights. The CCPA provides for fines and allows private litigants affected by certain data breaches to recover significant statutory damages. The CCPA and other comprehensive U.S. state privacy laws exempt some data processed in the context of clinical trials, but these developments may further complicate compliance efforts, and increase legal risk and compliance costs for us, the third parties with whom we work. Similar laws are being considered in several other states, as well as at the federal and local levels, and we expect more states to pass similar laws in the future. We may be or may become subject to new laws governing the privacy of consumer health data, including reproductive, sexual orientation, and gender identity privacy rights. For example, Washington’s My Health My Data Act (MHMD) broadly defines consumer health data, places restrictions on processing consumer health data (including imposing stringent requirements for consents), provides consumers certain rights with respect to their health data, and creates a private right of action to allow individuals to sue for violations of the law. Other states have passed, are considering, and may adopt similar laws.
Outside the United States, an increasing number of laws, regulations, and industry standards govern data privacy and security. For example, we are subject to the European Union’s General Data Protection Regulation (EU GDPR) and the United Kingdom’s GDPR (UK GDPR) (collectively, GDPR) by virtue of our sponsoring of clinical trials within
32
Table of Contents
the European Economic Area (EEA) or United Kingdom (UK). The EEA consists of the 27 member states of the EU plus Norway, Liechtenstein and Iceland and is subject to EU laws. The GDPR imposes strict requirements for processing personal data of individuals within the EEA or UK or in the context of our activities within the EEA or UK. For example, under the GDPR, companies may face temporary or definitive bans on data processing and other corrective actions; fines of up to 20 million Euros under the EU GDPR, 17.5 million pounds sterling under the UK GDPR or, in each case, 4% of annual global revenue, whichever is greater. In addition to fines, a breach of the GDPR may result in regulatory investigations, reputational damage, orders to cease/change our data processing activities, enforcement notices, and/or civil claims, including private litigation related to processing of personal data brought by classes of data subjects or consumer protection organizations authorized at law to represent their interests. In the EEA, the Network and Information Security Directive (NIS2) regulates resilience and incident response capabilities of entities operating in a number of sectors, including the health sector. Non-compliance with NIS2 may lead to administrative fines of a maximum of 10 million Euros or up to 2% of the total worldwide revenue of the preceding fiscal year.
In the ordinary course of business, we transfer personal data from the EU, UK and other jurisdictions to the United States or other countries. The EU, UK and other jurisdictions have enacted laws requiring data to be localized or limiting the transfer of personal data to other countries. In particular, the EU and other member states of the EEA and the UK have significantly restricted the transfer of personal data to the United States and other countries whose privacy laws it generally believes are inadequate. Other jurisdictions may adopt or have already adopted similarly stringent data localization and cross-border data transfer laws. Although there are currently various mechanisms that may be used to transfer personal data from the EEA and UK to the United States and other third countries in compliance with law, such as the EU standard contractual clauses, the UK’s International Data Transfer Agreement/Addendum, and the EU-U.S. Data Privacy Framework and the UK extension thereto (which allows for transfers to relevant U.S.-based organizations who self-certify compliance and participate in the Framework), these mechanisms are subject to legal challenges, and there is no assurance that we can satisfy or rely on these measures to lawfully transfer personal data to the United States and other third countries. If there is no lawful manner for us to transfer personal data from the EEA, the UK or other jurisdictions to the United States or other countries in which we operate, or if the requirements for a legally-compliant transfer are too onerous, we could face significant adverse consequences, including the interruption or degradation of our operations, the need to relocate part of or all of our business or data processing activities to other jurisdictions (such as Europe) at significant expense, increased exposure to regulatory actions, substantial fines and penalties, the inability to transfer data and work with partners, vendors and other third parties, and injunctions against our processing or transferring of personal data necessary to operate our business. Additionally, companies that transfer personal data out of the EEA and UK to other jurisdictions, particularly to the United States, are subject to increased scrutiny from regulators, individual litigants, and activist groups. Some European regulators have ordered certain companies to suspend or permanently cease certain transfers out of Europe for allegedly violating the GDPR’s cross-border data transfer limitations.
Additionally, the U.S. Department of Justice issued a rule entitled the Preventing Access to U.S. Sensitive Personal Data and Government-Related Data by Countries of Concern or Covered Persons, which places additional restriction on certain data transactions involving countries of concern (e.g., China, Russia, Iran) and covered persons (i.e., individuals and entities who are designated as such by the U.S. Attorney General or considered “foreign persons” and are majority-owned by, organized under the laws of, a primary resident in, or a contractor of, a covered person or country of concern, as applicable) that may impact certain business activities such as vendor engagements, sale or sharing of data, employment of certain individuals, and investor agreements. Violations of the rule could lead to significant civil and criminal fines and penalties. The rule applies regardless of whether data is anonymized, key-coded, pseudonymized, de-identified or encrypted, which presents particular challenges for companies like ours and may impact our ability to engage in transactions or agreements with certain third parties in the future.
In addition to data privacy and security laws, we are contractually subject to industry standards adopted by industry groups and, we may become subject to such obligations. We are also bound by contractual obligations related to data privacy and security, and our efforts to comply with such obligations may not be successful. We also publish privacy policies, marketing materials, whitepapers, and other statements, such as statements related to compliance with certain certifications or self-regulatory principles concerning data privacy, and security. Regulators in the United States are increasingly scrutinizing these statements, and if these policies, materials or statements are found
33
Table of Contents
to be deficient, lacking in transparency, deceptive, unfair, misleading, or misrepresentative of our practices, we may be subject to investigation, enforcement actions by regulators or other adverse consequences.
Our employees and personnel use generative artificial intelligence (AI) and/or automated decision-making technologies to perform their work, and the disclosure and use of personal data in AI technologies is subject to various privacy laws and other privacy obligations. Governments have passed and are likely to pass additional laws and regulations regulating AI and/or automated decision-making technologies. Our use of this technology could result in additional compliance costs, regulatory investigations and actions, and lawsuits. If we are unable to use AI and/or automated decision-making technologies, it could make our business less efficient and result in competitive disadvantages.
Obligations related to data privacy and security (and consumers’ data privacy expectations) are quickly changing, becoming increasingly stringent, and creating uncertainty. Additionally, these obligations may be subject to differing applications and interpretations, which may be inconsistent or conflict among jurisdictions. Preparing for and complying with these obligations requires us to devote significant resources, which may necessitate changes to our services, information technologies, systems, and practices and to those of any third parties that process personal data on our behalf. In addition, these obligations may require us to change our business model.
We may at times fail (or be perceived to have failed) in our efforts to comply with our data privacy and security obligations. Moreover, despite our efforts, our personnel or third parties with whom we work may fail to comply with such obligations, which could negatively impact our business operations. If we or the third parties with whom we work fail, or are perceived to have failed, to address or comply with applicable data privacy and security obligations, we could face significant consequences, including but not limited to: government enforcement actions (e.g., investigations, fines, penalties, audits, inspections, and similar); litigation (including class-action claims) and mass arbitration demands; additional reporting requirements and/or oversight; bans or restrictions on processing personal data; orders to destroy or not use personal data; and imprisonment of company officials. In particular, plaintiffs have become increasingly more active in bringing privacy-related claims against companies, including class claims and mass arbitration demands. Some of these claims allow for the recovery of statutory damages on a per violation basis, and, if viable, carry the potential for monumental statutory damages, depending on the volume of data and the number of violations. Any of these events could have a material adverse effect on our reputation, business, or financial condition, including but not limited to: interruptions or stoppages in our business operations (including, as relevant, clinical trials); inability to process personal data or to operate in certain jurisdictions; limited ability to develop or commercialize our products; expenditure of time and resources to defend any claim or inquiry; adverse publicity; or substantial changes to our business model or operations.
If we, or any contract manufacturers or suppliers we engage, fail to comply with environmental, health and safety laws and regulations, we could become subject to fines or penalties or incur costs that could harm our business.
We and our third-party contractors are subject to numerous federal, state, local and foreign environmental, health and safety laws and regulations, including those governing laboratory procedures and the handling, use, storage, treatment and disposal of hazardous materials and wastes. Our operations involve the use of hazardous and flammable materials, including chemicals and biological materials. Our operations also produce hazardous waste products. We generally contract with third parties for the disposal of these materials and wastes. We cannot eliminate the risk of contamination or injury from these materials. In the event of contamination or injury resulting from our use of hazardous materials, we could be held liable for any resulting damages, and any liability could exceed our resources, including any available insurance. We could also be held liable for unexpected safety events that could happen in our business offices.
In addition, our leasing and operation of real property may subject us to liability pursuant to certain of these laws or regulations. Under existing United States environmental laws and regulations, current or previous owners or operators of real property and entities that disposed or arranged for the disposal of hazardous substances may be held strictly, jointly and severally liable for the cost of investigating or remediating contamination caused by hazardous substance releases, even if they did not know of and were not responsible for the releases.
We could incur significant costs and liabilities which may adversely affect our financial condition and operating results for failure to comply with such laws and regulations, including, among other things, civil or criminal fines and
34
Table of Contents
penalties, property damage and personal injury claims, costs associated with upgrades to our facilities or changes to our operating procedures, or injunctions limiting or altering our operations.
Although we maintain liability insurance to cover us for costs and expenses that we may incur due to injuries to our employees, this insurance may not provide adequate coverage against potential liabilities. We do not maintain insurance for environmental liability or toxic tort claims that may be asserted against us in connection with our storage or disposal of biological, hazardous or radioactive materials.
In addition, we may incur substantial costs in order to comply with current or future environmental, health and safety laws and regulations. These current or future laws and regulations, which are becoming increasingly more stringent, may impair our research, development or production efforts. Our failure to comply with these laws and regulations also may result in substantial fines, penalties or other sanctions.
Our business entails a significant risk of product liability and our ability to obtain sufficient insurance coverage could have a material and adverse effect on our business, financial condition, results of operations, and prospects. If product liability lawsuits are brought against us, we may incur substantial liabilities and may be required to limit, delay or cease commercialization of our products.
When we conduct clinical trials of our current and any future product candidates, we may be exposed to significant product liability risks inherent in the development, testing, manufacturing and marketing of therapeutic treatments. Product liability claims could delay or prevent completion of our development programs. If we succeed in marketing products, if approved, such claims could result in an FDA investigation of the safety and effectiveness of our products, our manufacturing processes and facilities or our marketing programs and potentially a recall of our products or more serious enforcement action by U.S. or foreign regulatory authorities, limitations on the approved indications for which they may be used or suspension or withdrawal of approvals. If we cannot successfully defend ourselves against product liability claims, we may incur substantial liabilities or be required to limit, delay or cease the commercialization of our products. Regardless of the merits or eventual outcome, liability claims may also result in decreased demand for our products, termination of clinical trial sites or entire trial programs, withdrawal of clinical trial participants, injury to our reputation and significant negative media attention, significant costs to defend the related litigation, a diversion of management’s time and our resources from our business operations, substantial monetary awards to trial participants or patients, loss of revenue, the inability to commercialize any products that we may develop, and a decline in our stock price.
We currently maintain $5.0 million in general liability insurance and $5.0 million in product liability insurance. We may, however, need to obtain higher levels of insurance coverage for later stages of clinical development or marketing any of our product candidates. Any insurance we have or may obtain may not provide sufficient coverage against potential liabilities. Furthermore, clinical trial and product liability insurance is becoming increasingly expensive. As a result, we may be unable to obtain sufficient insurance at a reasonable cost to protect us against losses caused by product liability claims that could have a material and adverse effect on our business, financial condition, results of operations, and prospects. Our inability to obtain and retain sufficient product liability insurance at an acceptable cost to protect against potential product liability claims could prevent or inhibit the commercialization of our product candidates. Although we will maintain such insurance, any claim that may be brought against us could result in a court judgment or settlement in an amount that is not covered, in whole or in part, by our insurance or that is in excess of the limits of our insurance coverage. Our insurance policies will also have various exclusions, and we may be subject to a product liability claim for which we have no coverage. We may have to pay any amounts awarded by a court or negotiated in a settlement that exceed our coverage limitations or that are not covered by our insurance, and we may not have, or be able to obtain, sufficient capital to pay such amounts.
Our employees, independent contractors, consultants and vendors may engage in misconduct or other improper activities, including noncompliance with regulatory standards and requirements and insider trading.
We are exposed to the risk of employee fraud or other illegal activity by our employees, independent contractors, consultants and vendors. Misconduct by these parties could include intentional, reckless and/or negligent conduct that fails to comply with FDA regulations, provide true, complete and accurate information to the FDA or other comparable foreign regulatory authorities, comply with manufacturing standards we may establish, comply with healthcare fraud and abuse laws and regulations, report financial information or data accurately or disclose
35
Table of Contents
unauthorized activities to us. If we obtain FDA approval of any of our current or future product candidates and begin commercializing those products in the United States, our potential exposure under these laws will increase significantly, and our costs associated with compliance with these laws will likely increase. In particular, sales, marketing and business arrangements in the healthcare industry are subject to extensive laws and regulations intended to prevent fraud, kickbacks, self-dealing and other abusive practices. These laws and regulations may restrict or prohibit a wide range of pricing, discounting, marketing and promotion, sales commission, customer incentive programs and other business arrangements. Employee misconduct could also involve the improper use of information obtained in the course of clinical trials, which could result in regulatory sanctions and serious harm to our reputation. Additionally, we are subject to the risk that a person could allege such fraud or other misconduct, even if none occurred. It is not always possible to identify and deter employee misconduct, and the precautions we take to detect and prevent this activity may not be effective in controlling unknown or unmanaged risks or losses or in protecting us from governmental investigations or other actions or lawsuits stemming from a failure to comply with such laws or regulations. If any such actions are instituted against us, and we are not successful in defending ourselves or asserting our rights, those actions could have a material and adverse effect on our business, financial condition, results of operations, and prospects, including the imposition of significant civil, criminal and administrative penalties, damages, fines, disgorgement, imprisonment, the curtailment or restructuring of our operations, loss of eligibility to obtain approvals from the FDA or other comparable foreign regulatory authorities exclusion from participation in government contracting, healthcare reimbursement or other government programs, including Medicare and Medicaid, integrity oversight and reporting obligations, or reputational harm.
Our ability to use our net operating loss (NOL) carryforwards and certain other tax attributes to offset taxable income or taxes may be limited.
Under U.S. federal income tax law, federal NOLs incurred in tax years beginning after December 31, 2017 may be carried forward indefinitely, but the deductibility of such federal NOLs is limited to 80% of taxable income. In addition, under Sections 382 and 383 of the Internal Revenue Code of 1986, as amended (the Code), and corresponding provisions of state law, if a corporation undergoes an “ownership change,” which is generally defined as a greater than 50 percentage point change, by value, in its equity ownership over a three-year period, the corporation’s ability to utilize its pre-change NOL carryforwards and other pre-change tax attributes to offset its post-change income or taxes may be limited. We performed a detailed analysis as of December 31, 2025 and determined that we had not undergone an ownership change under Section 382 of the Code as of this date. In the future, if we undergo an ownership change, which may be outside of our control or occur as a result of this offering, our ability to utilize our NOL carryforwards and other tax attributes to offset post-change income or taxes may be limited. As a result, even if we attain profitability, our NOL carryforwards and other tax attributes may be subject to material usage limitations, which could negatively impact our future cash flows. In addition, for state income tax purposes, there may be periods during which the use of NOL carryforwards is suspended or otherwise limited, which could accelerate or permanently increase state taxes owed. For example, California imposed limits on the usability of California state NOL carryforwards to offset taxable income and certain business credits to offset California state tax liabilities in tax years beginning after 2023 and before 2027.
Changes and evolving requirements in tax laws or their interpretation could adversely affect our business.
The tax regimes we are subject to or operate under, including with respect to income and non-income taxes, are unsettled and may be subject to significant change. The One Big Beautiful Bill Act (the OBBBA) enacted in 2025, the Inflation Reduction Act (the IRA) enacted in 2022, the Coronavirus Aid, Relief, and Economic Security Act enacted in 2020, and the Tax Cuts and Jobs Act enacted in 2017 made many significant changes to the Code. Future guidance from the Internal Revenue Service and other tax authorities with respect to any legislation may affect us, and certain aspects of such legislation could be repealed or modified in future legislation or sunset in future years. It is possible that changes in or interpretations under the OBBBA or other tax legislation, or the enactment of new tax legislation, could increase our future tax liability, which could in turn adversely impact our business and future profitability. The U.S. government may enact further changes to the taxation of business entities including, among others, an increase in the corporate income tax rate, the imposition of minimum taxes or surtaxes on certain types of income and significant changes to the taxation of income derived from international operations. We are unable to predict what changes to the tax laws of the United States and other jurisdictions may be proposed or enacted in the future or what effect such changes would have on our business. Any of these or similar
36
Table of Contents
developments or changes to tax laws or rulings (which changes may have retroactive application) could result in adverse impacts to our financial condition, operating results and prospects and a material change to the tax considerations described herein.
Risks Related to Our Intellectual Property
If we or our future licensors are unable to obtain, maintain, defend, and enforce patent and other intellectual property protection for our current or future product candidates and technologies in the United States or other countries, or if the scope of the patent or other intellectual property protection obtained is not sufficiently broad, we may not be able to compete effectively.
Our success is dependent in part on our ability to obtain and maintain proprietary or intellectual property protection in the United States and other countries for our current product candidates or any future product candidates, as well as our core technologies, including our manufacturing know-how, and on our ability to operate without infringing, misappropriating or otherwise violating the proprietary rights of others. We strive to protect and enhance the proprietary technology, inventions and improvements that are commercially important to the development of our business by seeking, maintaining, defending and enforcing our intellectual property, whether developed internally or licensed from third parties. We also rely on trade secrets, know-how, continuing technological innovation and in-licensing opportunities to develop, strengthen and maintain our proprietary position in our field.
The patent position of biotechnology companies generally is highly uncertain, involves complex legal and factual questions and has in recent years been the subject of much litigation. The degree of patent protection we require to successfully compete in the marketplace may be unavailable or severely limited in some cases and may not adequately protect our rights or permit us to gain or keep any competitive advantage. We cannot provide any assurances that any of our current or future owned patent applications, or patent applications we may license in the future, will mature into issued patents, and cannot provide any assurances that any such patents, if issued, will include claims with a scope sufficient to protect our current and future product candidates or otherwise provide any competitive advantage. Additionally, patents can be enforced only in those jurisdictions in which the patent has issued. Furthermore, patents have a limited lifespan. In the United States, the natural expiration of a patent is generally 20 years after its first nonprovisional U.S. filing. The natural expiration of a patent outside of the United States varies in accordance with provisions of applicable local law, but is generally 20 years from the earliest local filing date. Various extensions may be available; however, the life of a patent, and the protection it affords, is limited. Given the amount of time required for the development, testing and regulatory review of new therapeutic candidates, patents protecting such candidates might expire before or shortly after such candidates are commercialized.
To the extent we develop a licensed patent portfolio, it may not provide us with adequate and continuing patent protection sufficient to exclude others from commercializing therapeutics similar to our product candidates, including biosimilar and interchangeable versions of such therapeutics. In addition, any patent portfolio that is licensed to us is, or may be, licensed to third parties outside our licensed field, and such third parties may have certain enforcement rights. Thus, patents licensed to us could be put at risk of being invalidated or interpreted narrowly or held unenforceable in litigation filed by or against another licensee or in administrative proceedings brought by or against another licensee in response to such litigation or for other reasons.
Other parties may have developed technologies that are related or competitive to our own and such parties may have filed or may file patent applications, or may have received or may receive patents, claiming inventions that may overlap or conflict with those claimed in our own current or future patent applications or issued patents. Publication of discoveries in the scientific literature lags behind the actual discoveries, and patent applications in the United States and in other jurisdictions are typically not published until 18 months after filing, or in some cases not at all. Therefore, we cannot know with certainty whether the inventors of our patents and applications were the first to make the inventions claimed in those patents or pending patent applications, or that we or our future licensors were the first to file for patent protection of such inventions. Further, we cannot assure that all of the potentially relevant prior art relating to our current and future patents and patent applications has been found. If such prior art exists, it can invalidate a patent or prevent a patent from issuing from a pending patent application. As a result, the issuance, inventorship, ownership, scope, validity and commercial value of our current and future patent rights cannot be
37
Table of Contents
predicted with any certainty. Further, if the breadth or strength of protection provided by our current and future patents and patent applications is threatened, regardless of the outcome, it could dissuade companies from collaborating with us to license, develop or commercialize current or future product candidates.
In addition, the patent prosecution process is expensive and time-consuming, and we may not be able to file and prosecute all necessary or desirable patent applications at a reasonable cost or in a timely manner. Moreover, the scope of the claims initially submitted for examination may be significantly narrowed by the time they issue, if at all. It is also possible that we will fail to identify patentable aspects of our research and development output before it is too late to obtain patent protection. We cannot provide any assurances that we will be able to pursue or obtain additional patent protection based on our research and development efforts, or that any such patents or other intellectual property we generate will provide any competitive advantage. Moreover, we may not have the right to control the preparation, filing and prosecution of patent applications, or to control the maintenance of the patents, covering technology that we may license from third parties in the future. Therefore, these patents and applications may not be filed, prosecuted or maintained in a manner consistent with the best interests of our business.
Even if we acquire patent protection that we expect should enable us to maintain competitive advantage, the issuance of a patent is not conclusive as to its inventorship, ownership, scope, validity or enforceability. Third parties, including former employees, consultants, collaborators and competitors, may challenge the inventorship, ownership, scope, validity or enforceability thereof, which may result in such patents being narrowed, invalidated or held unenforceable. We may become involved in opposition, reexamination, inter partes review, post-grant review, derivation, interference, or similar proceedings in the United States or abroad challenging the claims of our patents, once issued. Furthermore, patents may be challenged in court, once issued. Competitors may have filed patent applications before we did. A competitor may also claim that we are infringing its patents and that we therefore cannot practice our technology as claimed under our patent applications and patents, if issued. As a result, one or more claims of our current or future patents may be narrowed, invalidated or held unenforceable. In litigation, a competitor could claim that our current or future patents, if issued, are not valid for a number of reasons. If a court agrees, we would lose our rights to those challenged patents.
Even if they are unchallenged, our patents and pending patent applications, if issued, may not provide us with any meaningful protection or prevent competitors from designing around our patent claims to circumvent our patents by developing similar or alternative technologies or therapeutics in a non-infringing manner. For example, even if we have a valid and enforceable patent, we may not be able to exclude others from practicing our invention if the other party can show that they used the invention in commerce before our filing date or the other party benefits from an ex-U.S. compulsory license. If the patent protection provided by the patents and patent applications we hold or may pursue in the future with respect to our current and future product candidates is not sufficiently broad to impede such competition, our ability to successfully commercialize our product candidates could be negatively affected, which would harm our business, financial condition, results of operations, and prospects.
Certain regulatory exclusivities may be available. However, the scope of such regulatory exclusivities is subject to change, and may not provide us with adequate and continuing protection sufficient to exclude others from commercializing products similar to our product candidates.
We cannot ensure that patent rights relating to inventions described and claimed in our pending patent applications will issue or that patents based on our patent applications will not be challenged and rendered invalid and/or unenforceable.
The patent application process is subject to numerous risks and uncertainties, and there can be no assurance that we or any of our potential future collaborators will be successful in protecting our product candidates by obtaining, enforcing and defending patents. For example, we may not be aware of all third-party intellectual property rights potentially relating to our product candidates or their intended uses, and as a result the impact of such third-party intellectual property rights upon the patentability of our own patents and patent applications is highly uncertain. We have issued and pending U.S. and foreign patent applications in our portfolio covering aspects of our therapeutic programs; however, we cannot predict:
| ∎ | if and when patents may issue based on our patent applications; |
| ∎ | the scope of protection of any patent issuing based on our patent applications; |
38
Table of Contents
| ∎ | whether the claims of any patent issuing based on our patent applications will provide protection against competitors; |
| ∎ | whether or not third parties will find ways to invalidate or circumvent our patent rights; |
| ∎ | whether or not others will obtain patents claiming aspects similar to those covered by our patents and patent applications; |
| ∎ | whether we will need to initiate litigation or administrative proceedings to enforce and/or defend our patent rights which will be costly whether we win or lose; |
| ∎ | whether the patent applications that we own or in-license will result in issued patents with claims that cover our product candidates or uses thereof in the United States or in other foreign countries; and/or |
| ∎ | whether we may experience patent office interruption or delays to our ability to timely secure patent coverage to our product candidates. |
We cannot be certain that the claims in our pending patent applications directed to our current and future product candidates and/or technologies will be considered patentable by the United States Patent and Trademark Office (USPTO) or by patent offices in foreign countries. There can be no assurance that any such patent applications will issue as granted patents. One aspect of the determination of patentability of our inventions depends on the scope and content of the “prior art,” information that was or is deemed available to a person of skill in the relevant art prior to the priority date of the claimed invention. There may be prior art of which we are not aware that may affect the patentability of our patent claims or, if issued, affect the validity or enforceability of a patent claim. Even if the patents do issue based on our patent applications, third parties may challenge the validity, enforceability or scope thereof, which may result in such patents being narrowed, invalidated or held unenforceable. Furthermore, even if they are unchallenged, patents in our portfolio may not adequately exclude third parties from practicing relevant technology or prevent others from designing around our claims. If the breadth or strength of our intellectual property position with respect to our current or future product candidates is threatened, it could dissuade companies from collaborating with us to develop and threaten our ability to commercialize our product candidates.
Our pending patent applications may be challenged in the USPTO or in patent offices in foreign countries. Also, because the issuance of a patent is not conclusive as to its scope, validity or enforceability, even issued patents may later be found invalid or unenforceable or may be modified or revoked in proceedings instituted by third parties before various patent offices or in courts. For example, our pending patent applications may be subject to third-party pre-issuance submissions of prior art to the USPTO or patent offices in foreign countries or our issued patents may be subject to post-grant review, oppositions, derivations, reexaminations or inter partes review proceedings in the United States or elsewhere, challenging our patent rights or the patent rights of others. An adverse determination in any such challenges may result in loss of exclusivity or in our patent claims being narrowed, invalidated, or held unenforceable, in whole or in part, which could limit our ability to stop others from using or commercializing similar or identical technologies and therapeutics, or limit the duration of the patent protection of our technologies and product candidates. In the event of litigation or administrative proceedings, we cannot be certain that the claims in any of our issued patents will be considered valid by courts or administrative authorities in the United States or foreign countries. In addition, given the amount of time required for the development, testing and regulatory review of new product candidates, patents protecting such candidates might expire before or shortly after such candidates are commercialized. As a result, only limited protection may be available and our patent portfolio may not provide us with sufficient rights or permit us to gain or keep any competitive advantage. Any failure to obtain or maintain patent protection with respect to our product candidates or their uses could have a material adverse effect on our business, financial condition, results of operations, and prospects.
We may not be able to protect our intellectual property rights throughout the world.
Although we have pending patent applications in the United States and other countries, filing, prosecuting, maintaining, enforcing and defending patents in all countries throughout the world would be prohibitively expensive, and our intellectual property rights in some countries outside the United States can be less extensive than those in the United States. In addition, the laws of some foreign countries do not protect intellectual property rights to the same extent as federal and state laws in the United States. Consequently, we may not be able to prevent third parties from practicing our inventions in all countries outside the United States or from selling or importing products made using our inventions in and into the United States or other jurisdictions. Competitors may use our technologies
39
Table of Contents
in jurisdictions where we have not obtained patent protection to develop their own therapeutics and, further, may export otherwise infringing therapeutics to territories where we have patent protection, but enforcement is not as strong as that in the United States. These therapeutics may compete with our product candidates, and our patents, the patents of our future licensors or other intellectual property rights may not be effective or sufficient to prevent them from competing.
Many companies have encountered significant problems in protecting and defending intellectual property rights in foreign jurisdictions. The legal systems of many foreign countries do not favor the enforcement of patents, trade secrets and other intellectual property protection, particularly those relating to biopharmaceutical products, which could make it difficult in those jurisdictions for us to stop the infringement or misappropriation of our patents or other intellectual property rights, or the marketing of competing therapeutics in violation of our proprietary rights. Proceedings to enforce our patent and other intellectual property rights in foreign jurisdictions could result in substantial costs and divert our efforts and attention from other aspects of our business, could put our patents or the patents of our licensors at risk of being invalidated or interpreted narrowly and our patent applications or the patent applications of our licensors at risk of not issuing and could provoke third parties to assert claims of infringement or misappropriation against us. We may not prevail in any lawsuits that we initiate, and the damages or other remedies awarded, if any, may not be commercially meaningful. Accordingly, our efforts to enforce our intellectual property rights around the world may be inadequate to obtain a significant commercial advantage from the intellectual property that we develop or may license.
In addition, geopolitical actions in the United States and in foreign countries could increase the uncertainties and costs surrounding the prosecution or maintenance of our patent applications and the maintenance, enforcement or defense of our issued patents.
Many countries, including China and India, have compulsory licensing laws under which a patent owner may be compelled to grant licenses to third parties. In addition, many countries limit the enforceability of patents against government agencies or government contractors. In these countries, we or any of our future licensors may have limited remedies if patents are infringed or if we or our licensors are compelled to grant a license to a third party, which could materially diminish the value of such patent. If we are forced to grant a license to third parties with respect to any patents relevant to our business, our competitive position may be impaired, and our business, financial condition, results of operations, and prospects may be adversely affected.
Furthermore, we may not be able to prevent, alone or with our future licensors, misappropriation of our intellectual property rights, particularly in countries where the laws may not protect those rights as fully as in the United States. Any litigation or other proceedings to enforce our intellectual property rights may fail, and even if successful, may result in substantial costs and distract our management and other employees.
We may not identify relevant third-party patents or may incorrectly interpret the relevance, scope or expiration of a third-party patent, which might adversely affect our ability to develop and market our product candidates.
As the biotechnology industry expands and more patents are issued, the risk increases that our product candidates may be subject to claims of infringement of the patent rights of third parties. There can be no assurance that our operations do not, or will not in the future, infringe existing or future third-party patents. Identification of third-party patent rights that may be relevant to our operations is difficult because patent searching is imperfect due to differences in terminology among patents, incomplete databases and the difficulty in assessing the meaning of patent claims. We cannot guarantee that any of our patent searches or analyses, including the identification of relevant patents, the scope of patent claims or the expiration of relevant patents, are complete or thorough, nor can we be certain that we have identified each and every third-party patent and pending application in the United States and abroad that is relevant to or necessary for the commercialization of our current or future product candidates in any jurisdiction.
Numerous U.S. and foreign patents and pending patent applications exist in our market that are owned by third parties. Our competitors in both the United States and abroad, many of which have substantially greater resources and have made substantial investments in patent portfolios and competing technologies, may have applied for or obtained or may in the future apply for and obtain, patents that will prevent, limit or otherwise interfere with our ability to make, use and sell our product candidates. We do not always conduct independent reviews of pending
40
Table of Contents
patent applications of, and patents issued to, third parties. Patent applications in the United States and elsewhere are typically published approximately 18 months after the earliest filing for which priority is claimed, with such earliest filing date being commonly referred to as the priority date. Certain U.S. applications that will not be filed outside the United States can remain confidential until patents issue. In addition, patent applications in the United States and elsewhere can be pending for many years before issuance, or unintentionally abandoned patents or applications can be revived. Furthermore, pending patent applications that have been published can, subject to certain limitations, be later amended in a manner that could cover our current or future technologies, our product candidates or the use of our product candidates. As such, there may be patent applications now pending or recently revived patents of which we are unaware. These patent applications may later result in issued patents, or the revival of previously abandoned patents, that will prevent, limit or otherwise interfere with our ability to make, use or sell our product candidates.
The scope of a patent claim is determined by the literal language of the patent claim, an interpretation of the law, the written disclosure in a patent and the patent’s prosecution history. Our interpretation of the relevance or the scope of a patent or a pending application may be incorrect. For example, we may incorrectly determine that our current or future product candidates are not covered by a third-party patent or may incorrectly predict whether a third party’s pending application will issue with claims of relevant scope. Alternatively, we may incorrectly determine that the Hatch-Waxman Amendments are a defense for a safe harbor to infringement of a patent we consider relevant to the research or clinical development of any of our current or future product candidates. Our determination of the expiration date of any patent in the United States or abroad that we consider relevant may be incorrect, and we may incorrectly conclude that a third-party patent is invalid and unenforceable or not infringed. Our failure to identify and correctly interpret relevant patents may negatively impact our ability to develop and market our current and future product candidates.
We cannot provide any assurances that third-party patents do not exist which might be enforced against our current technology, including our research programs, product candidates, their respective methods of use, manufacture and formulations thereof, and could result in either an injunction prohibiting our manufacture or future sales, or, with respect to our future sales, an obligation on our part to pay royalties and/or other forms of compensation to third parties, which could be significant.
If we are sued for infringing intellectual property rights of third parties, such litigation could be costly and time consuming and could prevent or delay us from developing or commercializing our current and future product candidates.
Our commercial success depends, in part, on our ability to develop, manufacture, market and sell our current and future product candidates without infringing the intellectual property and other proprietary rights of third parties. Third parties may allege that we have infringed or misappropriated their intellectual property. Litigation or other legal proceedings relating to intellectual property claims, with or without merit, is unpredictable and generally expensive and time consuming and, even if resolved in our favor, is likely to divert significant resources from our core business, including distracting our technical and management personnel from their normal responsibilities. In addition, there could be public announcements of the results of hearings, motions or other interim proceedings or developments and if securities analysts or investors perceive these results to be negative, it could have a substantial adverse effect on the market price of our common stock. Such litigation or proceedings could substantially increase our operating losses and reduce the resources available for development activities or any future sales, marketing or distribution activities. We may not have sufficient financial or other resources to adequately conduct such litigation or proceedings. Some of our competitors may be able to sustain the costs of such litigation or proceedings more effectively than we can because of their greater financial resources and more mature and developed intellectual property portfolios. Uncertainties resulting from the initiation and continuation of patent litigation or other proceedings could have a material adverse effect on our ability to compete in the marketplace.
There is a substantial amount of intellectual property litigation in the biotechnology industries, and we may become party to, or threatened with, litigation or other adversarial proceedings regarding intellectual property rights with respect to our product candidates. We cannot be certain that our product candidates and other proprietary technologies we may develop will not infringe existing or future patents owned by third parties. Third parties may assert infringement claims against us based on existing or future intellectual property rights. If we are found to
41
Table of Contents
infringe a third party’s intellectual property rights, we could be forced, including by court order, to cease developing, manufacturing or commercializing the infringing product candidate or product. Alternatively, we may be required to obtain a license from such third party in order to use the infringing technology and continue developing, manufacturing or marketing the infringing product candidate or product. However, we may not be able to obtain any required license on commercially reasonable terms or at all. Even if we were able to obtain a license, it could be non-exclusive, thereby giving our competitors access to the same technologies licensed to us. In addition, we could be found liable for monetary damages, including treble damages and attorneys’ fees if we are found to have willfully infringed a patent. A finding of infringement could prevent us from commercializing our product candidates or force us to cease some of our business operations, which could materially harm our business, financial condition, results of operations, and prospects. Claims that we have misappropriated the confidential information or trade secrets of third parties could have a similar negative impact on our business.
We may not be aware of patents that have already been issued and that a third party, for example, a competitor in the fields in which we are developing our product candidates, might assert are infringed by our current or future product candidates, including claims to compositions, formulations, methods of manufacture or methods of use or treatment that cover such therapeutic candidates. It is also possible that patents owned by third parties of which we are aware, but which we do not believe are relevant to our product candidates and other proprietary technologies we may develop, could be found to be infringed by our current or future product candidates. In addition, because patent applications can take many years to issue, there may be currently pending patent applications that may later result in issued patents that our current or future product candidates may infringe. Our competitors, many of which have substantially greater resources and have made substantial investments in patent portfolios and competing technologies, may have applied for or obtained or may in the future apply for and obtain, patents that will prevent, limit or otherwise interfere with our ability to make, use and sell our current or future product candidates. The pharmaceutical and biotechnology industries have produced a considerable number of patents, and it may not always be clear to industry participants, including us, which patents cover various types of products or methods of use. The coverage of patents is subject to interpretation by the courts, and the interpretation is not always uniform. If we were sued for patent infringement, we would need to demonstrate that our product candidates, proprietary technologies or methods either do not infringe the patent claims of the relevant patent or that the patent claims are invalid or unenforceable, and we may not be able to do this. Proving invalidity may be difficult. In order to successfully challenge the validity of any such U.S. patent in federal court, we would need to overcome a presumption of validity. As this burden is a high one requiring us to present clear and convincing evidence as to the invalidity of any such U.S. patent claim, a court of competent jurisdiction may not invalidate the claims of any such U.S. patent. Even if we are successful in these proceedings, we may incur substantial costs and the time and attention of our management and scientific personnel could be diverted in pursuing these proceedings, which could have a material adverse effect on our business and operations. Furthermore, because of the substantial amount of discovery required in connection with intellectual property litigation, there is a risk that some of our confidential information could be compromised by disclosure during litigation. In addition, we may not have sufficient resources to bring these actions to a successful conclusion.
We may choose to challenge the enforceability or validity of claims in a third party’s patent by requesting that the USPTO review the patent claims in an ex-parte re-exam, inter partes review or post-grant review proceedings. These proceedings are expensive and may consume our time or other resources. We may choose to challenge a third party’s patent in patent opposition proceedings in foreign patent offices as well. The costs of these opposition proceedings could be substantial, and may consume our time or other resources. If we fail to obtain a favorable result at the USPTO or other patent office then we may be exposed to litigation by a third party alleging that the patent may be infringed by our product candidates or proprietary technologies.
We may become involved in lawsuits to protect or enforce our patents or other intellectual property, which could be expensive, time consuming and unsuccessful.
Competitors or other third parties may infringe or otherwise violate our current and future patents, trademarks or other intellectual property. To counter infringement or unauthorized use, we may be required to file infringement claims, which can be expensive and time consuming and divert the time and attention of our management and scientific personnel. Our pending patent applications cannot be enforced against third parties practicing the technology claimed in such applications unless and until a patent issues from such applications. Any claims we assert against perceived infringers could provoke these parties to assert counterclaims against us alleging that we
42
Table of Contents
infringe their patents, in addition to counterclaims asserting that our patents are invalid or unenforceable, or both. In patent litigation in the United States, defendant counterclaims alleging invalidity and/or unenforceability are commonplace. Grounds for a validity challenge could be an alleged failure to meet any of several statutory requirements, including lack of novelty, obviousness, non-enablement or insufficient written description. Grounds for an unenforceability assertion could be an allegation that someone connected with prosecution of the patent withheld relevant information from the USPTO or made a misleading statement during prosecution. Third parties may also raise similar validity claims before the USPTO in post-grant proceedings such as ex parte reexaminations, inter partes review, or post-grant review, or oppositions or similar proceedings outside the United States, in parallel with litigation or even outside the context of litigation. For the patents and patent applications that we may license in the future, we may have limited or no right to participate in the defense of any licensed patents against challenge by a third party. If a defendant were to prevail on a legal assertion of invalidity or unenforceability, we could lose at least part, and perhaps all, of any future patent protection on our current or future product candidates. Such a loss of patent protection could harm our business. The outcome following legal assertions of invalidity and unenforceability is unpredictable. In any patent infringement proceeding, there is a risk that a court will decide that a patent of ours is invalid or unenforceable, in whole or in part, and that we do not have the right to stop the other party from using the invention at issue. There is also a risk that, even if the validity of such patents is upheld, the court will construe the patent’s claims narrowly or decide that we do not have the right to stop the other party from using the invention at issue on the grounds that our patent claims do not cover the invention, or decide that the other party’s use of our current or future patented technology falls under the safe harbor to patent infringement under 35 U.S.C. §271(e)(1). An adverse outcome in a litigation or proceeding involving our patents could limit our ability to assert our patents against those parties or other competitors and may curtail or preclude our ability to exclude third parties from making and selling similar or competitive products. Any of these occurrences could adversely affect our competitive business position, business prospects and financial condition. Similarly, if we assert trademark infringement claims, a court may determine that the marks we have asserted are invalid or unenforceable, or that the party against whom we have asserted trademark infringement has superior rights to the marks in question. In such case, we could ultimately be forced to cease use of such trademarks. In any intellectual property litigation, even if we are successful, any award of monetary damages or other remedy we receive may not be commercially valuable.
Even if we establish infringement, the court may decide not to grant an injunction against further infringing activity and instead award only monetary damages, which may or may not be an adequate remedy. Furthermore, because of the substantial amount of discovery required in connection with intellectual property litigation, there is a risk that some of our confidential information could be compromised by disclosure during litigation. There could also be public announcements of the results of hearings, motions or other interim proceedings or developments. If securities analysts or investors perceive these results to be negative, it could have a material adverse effect on the price of shares of our common stock. Moreover, we cannot assure you that we will have sufficient financial or other resources to file and pursue such infringement claims, which typically last for years before they are concluded. Even if we ultimately prevail in such claims, the monetary cost of such litigation and the diversion of the attention of our management and scientific personnel could outweigh any benefit we receive as a result of the proceedings.
Further, interference or derivation proceedings provoked by third parties or brought by the USPTO or patent offices in foreign countries may be necessary to determine the priority of inventions with respect to, or the correct inventorship of, our patents or patent applications or those of our licensors. An unfavorable outcome could result in a loss of our current patent rights and could require us to cease using the related technologies or to attempt to license rights to it from the prevailing party. Our business could be harmed if the prevailing party does not offer us a license on commercially reasonable terms. Litigation, interference, derivation or other proceedings may result in a decision adverse to our interests and, even if we are successful, may result in substantial costs and distract our management and other employees.
Because of the expense and uncertainty of litigation, we may not be in a position to enforce our intellectual property rights against third parties.
Because of the expense and uncertainty of litigation, we may conclude that even if a third party is infringing our issued patents, any patents that may be issued as a result of our pending or future patent applications or other intellectual property rights, the risk-adjusted cost of bringing and enforcing such a claim or action may be too high or not in the best interest of our company or our stockholders, or it may be otherwise impractical or undesirable to enforce our intellectual
43
Table of Contents
property against some third parties. Our competitors or other third parties may be able to sustain the costs of complex patent litigation or proceedings more effectively than we can because of their greater financial resources and more mature and developed intellectual property portfolios. In such cases, we may decide that the more prudent course of action is to simply monitor the situation or initiate or seek some other non-litigious action or solution. In addition, the uncertainties associated with litigation could compromise our ability to raise the funds necessary to continue our clinical trials, continue our internal research programs, in-license needed technology or other product candidates, or enter into development partnerships that would help us bring our product candidates to market.
We may not be successful in obtaining or maintaining necessary rights to our product candidates through acquisitions and in-licenses.
Because our development programs have required, and may in the future require, the use of proprietary rights held by third parties, the growth of our business may depend in part on our ability to acquire, in-license or use these third-party proprietary rights. We may be unable to acquire or in-license any compositions, methods of use, processes or other third-party intellectual property rights from third parties that we identify as necessary for our product candidates on terms satisfactory to us or at all. For example, we are party to certain agreements with third parties, including a mouse platform services agreement, a non-exclusive license, a master services agreement and manufacturing agreement, pursuant to which we are obligated to make certain payments, including milestone payments and low single-digit royalties, in exchange for the right to use their respective technologies. If we fail to comply with our payment obligations or otherwise breach any of these agreements, we could lose our rights to use such third-party technologies or pay penalties, which could have a material adverse effect on our business, financial condition, results of operations and prospects. The licensing and acquisition of third-party intellectual property rights is a competitive area, and a number of more established companies may pursue strategies to license or acquire third-party intellectual property rights that we may consider attractive or necessary. These established companies may have a competitive advantage over us due to their size, capital resources and greater clinical development and commercialization capabilities. In addition, companies that perceive us to be a competitor may be unwilling to assign or license rights to us. We also may be unable to license or acquire third-party intellectual property rights on terms that would allow us to make an appropriate return on our investment or at all. If we are unable to successfully obtain rights to required third-party intellectual property rights, we may have to abandon development of the relevant program or product candidates, which could have a material adverse effect on our business, financial condition, results of operations, and prospects.
While we normally seek to obtain the right to control the prosecution of patent applications, and the maintenance and enforcement of the patents relating to our current and future product candidates, there may be times when the filing and prosecution activities for patents and patent applications relating to our current and future product candidates are controlled by our future licensors or collaboration partners. If any of our future licensors or collaboration partners fail to prosecute, maintain and enforce such patents and patent applications in a manner consistent with the best interests of our business, including by payment of all applicable fees for patents covering such product candidates, we could lose our rights to the intellectual property or our exclusivity with respect to those rights, our ability to develop and commercialize those product candidates may be adversely affected and we may not be able to prevent competitors from making, using and selling competing therapeutics. In addition, even where we have the right to control patent prosecution of patents and patent applications we have licensed to and from third parties, we may still be adversely affected or prejudiced by actions or inactions of our licensees, our future licensors and their counsel that took place prior to the date upon which we assumed control over patent prosecution.
We may enter into license agreements in the future with others to advance our existing or future research or allow commercialization of our existing or future product candidates. These licenses may not provide exclusive rights to use such intellectual property and technology in all relevant fields of use and in all territories in which we may wish to develop or commercialize our technology and therapeutics in the future.
Our future licensors may rely on third-party consultants or collaborators or on funds from third parties such that our future licensors are not the sole and exclusive owners of the patents we in-license. If other third parties have ownership rights to our future in-licensed patents, they may be able to license such patents to our competitors, and our competitors could market competing therapeutics and technology. This could have a material adverse effect on our competitive position, business, financial conditions, results of operations, and prospects.
44
Table of Contents
It is possible that we may be unable to obtain licenses at a reasonable cost or on reasonable terms, if at all. Even if we are able to obtain a license, it may be non-exclusive, thereby giving our competitors access to the same technologies licensed to us. In that event, we may be required to expend significant time and resources to redesign our technology, product candidates, or the methods for manufacturing them or to develop or license replacement technology, all of which may not be feasible on a technical or commercial basis. If we are unable to do so, we may be unable to develop or commercialize the affected product candidates, which could harm our business, financial condition, results of operations, and prospects significantly. We cannot provide any assurances that third-party patents do not exist which might be enforced against our current technology, manufacturing methods, product candidates or future methods or products resulting in either an injunction prohibiting our manufacture or future sales, or, with respect to our future sales, an obligation on our part to pay royalties and/or other forms of compensation to third parties, which could be significant.
Disputes may arise between us and our future licensors regarding intellectual property subject to a license agreement. In addition, the agreements under which we license intellectual property or technology from third parties are complex, and certain provisions in such agreements may be susceptible to multiple interpretations. The resolution of any contract interpretation disagreement that may arise could narrow what we believe to be the scope of our rights to the relevant intellectual property or technology, or increase what we believe to be our financial or other obligations under the relevant agreement, either of which could have a material adverse effect on our business, financial condition, results of operations, and prospects. Moreover, if disputes over intellectual property that we license in the future prevent or impair our ability to maintain our licensing arrangements on commercially acceptable terms, we may be unable to successfully develop and commercialize the affected product candidates, which could have a material adverse effect on our business, financial conditions, results of operations, and prospects.
In spite of our best efforts, our future licensors might conclude that we materially breached our license agreements and might therefore terminate the license agreements, thereby removing our ability to develop and commercialize product candidates and technology covered by these license agreements. If these in-licenses are terminated, or if the underlying patents fail to provide the intended exclusivity, competitors would have the freedom to seek regulatory approval of, and to market, therapeutics identical to ours. This could have a material adverse effect on our competitive position, business, financial conditions, results of operations, and prospects.
Any future collaborations that we enter into may not be successful. The success of our collaboration arrangements will depend heavily on the efforts and activities of our collaborators.
Our future licensors may retain certain rights under the relevant agreements with us, including the right to use the underlying technology for noncommercial academic and research use, to publish general scientific findings from research related to the technology, and to make customary scientific and scholarly disclosures of information relating to the technology. It is difficult to monitor whether our future licensors limit their use of the technology to these uses, and we could incur substantial expenses to enforce our rights to our licensed technology in the event of misuse.
In addition, the United States federal government retains certain rights in inventions produced with its financial assistance under the Patent and Trademark Law Amendments Act (Bayh-Dole Act). The federal government retains a “nonexclusive, nontransferable, irrevocable, paid-up license” for its own benefit. The Bayh-Dole Act also provides federal agencies with “march-in rights.” March-in rights allow the government, in specified circumstances, to require the contractor or successors in title to the patent to grant a “nonexclusive, partially exclusive, or exclusive license” to a “responsible applicant or applicants.” If the patent owner refuses to do so, the government may grant the license itself. We sometimes collaborate with academic institutions to accelerate our preclinical research or development. While it is our policy to avoid engaging university partners in projects in which there is a risk that federal funds may be commingled, we cannot be sure that any co-developed intellectual property will be free from government rights pursuant to the Bayh-Dole Act. If, in the future, we co-own or license in technology which is critical to our business that is developed in whole or in part with federal funds subject to the Bayh-Dole Act, our ability to enforce or otherwise exploit patents covering such technology may be adversely affected.
We may be subject to claims that we have wrongfully hired an employee from a competitor or that our employees, consultants or independent contractors have wrongfully used or disclosed confidential information of third parties.
As is common in the biotechnology industry, in addition to our employees, we engage the services of consultants to assist us in the development of our current and future product candidates. Many of these consultants, and many of
45
Table of Contents
our employees, were previously employed at, or may have previously provided or may be currently providing consulting services to, other biotechnology or pharmaceutical companies including our competitors or potential competitors. We could in the future be subject to claims that we or these individuals have inadvertently or otherwise used or disclosed intellectual property, trade secrets or other confidential information of any such individual’s current or former employers or clients. Although we try to ensure that our employees and consultants do not use the intellectual property, proprietary information, know-how or trade secrets of others in their work for us, we may become subject to claims that we caused an employee to breach the terms of his or her non-competition or non-solicitation agreement, or that we or these individuals have, inadvertently or otherwise, used or disclosed the alleged trade secrets or other proprietary information of a former employer or competitor.
While we may litigate to defend ourselves against these claims, even if we are successful, litigation could result in substantial costs and could be a distraction to management. If our defenses to these claims fail, in addition to requiring us to pay monetary damages, a court could prohibit us from using technologies or features that are essential to our current or future product candidates, if such technologies or features are found to incorporate or be derived from the trade secrets or other proprietary information of others. Moreover, any such litigation or the threat thereof may adversely affect our reputation, our ability to form strategic alliances or sublicense our rights to collaborators, engage with scientific advisors or hire employees or consultants, each of which would have an adverse effect on our business, financial condition, results of operations, and prospects. Even if we are successful in defending against such claims, litigation could result in substantial costs and be a distraction to management.
Changes in U.S. patent and ex-U.S. patent laws could diminish the value of patents in general, thereby impairing our ability to protect our current or future product candidates.
Changes in either the patent laws or interpretation of the patent laws in the United States or in other jurisdictions could increase the uncertainties and costs surrounding the prosecution of patent applications and the enforcement or defense of issued patents. In the United States, numerous recent changes to the patent laws and proposed changes to the rules of the USPTO may have a significant impact on our ability to protect our technology and enforce our intellectual property rights.
For example, the Leahy-Smith Act includes a number of significant changes to United States patent law. These changes include provisions that affect the way patent applications are prosecuted, redefine prior art, provide more efficient and cost-effective avenues for competitors to challenge the validity of patents and enable third-party submission of prior art to the USPTO during patent prosecution and additional procedures to attack the validity of a patent at USPTO-administered post-grant proceedings, including post-grant review, inter partes review and derivation proceedings. Because of a lower evidentiary standard in USPTO proceedings compared to the evidentiary standard in United States federal courts necessary to invalidate a patent claim, a third party could potentially provide evidence in a USPTO proceeding sufficient for the USPTO to hold a claim invalid even though the same evidence would be insufficient to invalidate the claim if first presented in a district court action. Accordingly, a third party may attempt to use the USPTO procedures to invalidate our patent claims that would not have been invalidated if first challenged by the third party as a defendant in a district court action.
Assuming that other requirements for patentability are met, and after March 2013, under the Leahy-Smith Act, the United States transitioned from a “first-to-invent” system to a “first-to-file” system in which, the first inventor to file a patent application will be entitled to the patent on an invention regardless of whether a third party was the first to invent the claimed invention. A third party that files a patent application in the USPTO after March 2013, but before we file an application covering the same invention, could therefore be awarded a patent covering an invention of ours even if we had made the invention before it was made by such third party. This will require us to be cognizant going forward of the time from invention to filing of a patent application. Since patent applications in the United States and most other countries are confidential for a period of time after filing or until issuance, we cannot be certain that we or our future licensors were the first to either (i) file any patent application related to our current or future product candidates and other technologies we may develop or (ii) invent any of the inventions claimed in our or our future licensor’s patents or patent applications. Even where we have a valid and enforceable patent, we may not be able to exclude others from practicing the claimed invention where the other party can show that they used the invention in commerce before our filing date or the other party benefits from a compulsory license. As such, Leahy-Smith Act and its implementation could increase the uncertainties and costs surrounding the prosecution of our
46
Table of Contents
patent applications and the enforcement or defense of our issued patents, all of which could have a material adverse effect on our business, financial condition, results of operations, and prospects.
In addition, the patent positions of companies in the development and commercialization of pharmaceuticals and biologics are particularly uncertain. Recent U.S. Supreme Court rulings have narrowed the scope of patent protection available in certain circumstances and weakened the rights of patent owners in certain situations. For example, in Amgen Inc. v. Sanofi, 598 U.S. 594, the U.S. Supreme Court held that claims with functional language may face high hurdles in fulfilling the enablement requirement. In addition, in Juno Therapeutics, Inc. v. Kite Pharma, Inc. 10 F.4th 1330 (Fed. Cir. 2021), the Federal Circuit held claims with functional language supported by few examples invalid for lack of written description. Recent Federal Circuit decisions, such as In re Cellect, LLC, 81 F.4th 1216 (Fed. Cir. 2023), raise questions regarding the award of patent term adjustment (PTA) for patents where related patents have been issued without a PTA. Thus, it cannot be said with certainty how PTA will or will not be viewed in the future and whether patent expiration dates may be impacted. We cannot predict how this and future decisions by the courts will affect our business. This combination of events has created uncertainty with respect to the validity and enforceability of patents once obtained. Depending on future actions by the U.S. Congress, the federal courts and the USPTO, the laws and regulations governing patents could change in unpredictable ways that could have a material adverse effect on our patent rights and our ability to protect, defend, and enforce our patent rights in the future. Any similar adverse changes in the patent laws of other jurisdictions could also have a material adverse effect on our business, financial condition, results of operations, and prospects.
Furthermore, in Europe, a new unitary patent system took effect June 1, 2023, which will significantly impact European patents, including those granted before the introduction of such a system. Under the unitary patent system, European applications have the option, upon grant of a patent, of becoming a Unitary Patent which will be subject to the jurisdiction of the Unitary Patent Court (UPC). As a result, all European patents, including those issued before the implementation of the UPC, now by default automatically fall under the jurisdiction of the UPC. It is uncertain how the UPC will impact granted European patents in the biotechnology and pharmaceutical industries. Our European patent applications, if issued, could be challenged in the UPC. During the first seven years of the UPC’s existence, the UPC legislation allows a patent owner to opt its European patents out of the jurisdiction of the UPC. We may decide to opt out our future European patents from the UPC, but doing so may preclude us from realizing the benefits of the UPC. Moreover, if we do not meet all of the formalities and requirements for opt-out under the UPC, our future European patents could remain under the jurisdiction of the UPC. The UPC will provide our competitors with a new forum to centrally revoke our European patents, and allow for the possibility of a competitor to obtain pan-European injunction. Such a loss of patent protection could have a material adverse impact on our business and our ability to commercialize our technology and product candidates and, resultantly, on our business, financial condition, prospects and results of operations.
Obtaining and maintaining our patent protection is dependent on compliance with various procedural, document submission, fee payment and other requirements imposed by governmental patent agencies, and our patent protection could be reduced or eliminated for non-compliance with these requirements.
Periodic maintenance fees, renewal fees, annuity fees and various other governmental fees on patents and/or applications will be due to be paid to the USPTO and various governmental patent agencies outside of the United States in several stages over the lifetime of the patents and/or applications. We employ an outside firm to pay these fees due to the USPTO and non-U.S. governmental patent agencies. The USPTO and various non-U.S. governmental patent agencies require compliance with a number of procedural, documentary, fee payment and other similar provisions during the patent application process. In the future, we may also depend on our licensors to take the necessary action to comply with these requirements with respect to our licensed intellectual property. We employ reputable law firms and other professionals to help us comply, and in many cases an inadvertent lapse can be cured by payment of a late fee or by other means in accordance with the applicable rules. However, there are situations in which non-compliance can result in abandonment or lapse of the patent or patent application, resulting in partial or complete loss of patent rights in the relevant jurisdiction. In such an event, our competitors might be able to enter the market and this circumstance would have a material adverse effect on our business.
The USPTO and various non-U.S. government agencies require compliance with certain foreign filing requirements during the patent application process. For example, in some countries, including the United States, China, India and
47
Table of Contents
some European countries, a foreign filing license is required before certain patent applications are filed. The foreign filing license requirements vary by country and depend on various factors, including where the inventive activity occurred, citizenship status of the inventors, the residency of the inventors and the invention owner, the place of business for the invention owner and the nature of the subject matter to be disclosed (e.g., items related to national security or national defense). In some, but not all cases, for example in China and India, a foreign filing license cannot be obtained retroactively in accordance with the applicable rules. There are situations, however, in which non-compliance can result in abandonment of a pending patent application or can be grounds for revoking or invalidating an issued patent in that jurisdiction, resulting in the loss of patent rights in the relevant jurisdiction. Other non-compliance events that could result in abandonment or lapse of a patent or patent application include, but are not limited to, failure to respond to official actions within prescribed time limits, non-payment of fees and failure to properly legalize and submit formal documents. In such an event, potential competitors might be able to enter the relevant markets with similar or identical products or technology, which could have a material adverse effect on our business, financial condition, results of operations, and prospects. In the future, we may also be dependent on licensors to take the necessary actions to comply with these requirements with respect to our in-licensed intellectual property.
If we are unable to protect the confidentiality of our trade secrets, our business and competitive position could be harmed.
In addition to seeking patent protection for our current and future product candidates, we rely on the protection of our trade secrets, unpatented know-how, technology and other proprietary information to maintain our competitive position. Although we have taken steps to protect our trade secrets and unpatented know-how, including entering into confidentiality agreements with third parties, and confidential information and inventions agreements with employees, consultants and advisors, we cannot guarantee that we have entered into applicable agreements with each party that may have or have had access to our trade secrets or proprietary information. In addition, we cannot provide any assurances that any party thereto will not breach the agreement and disclose our proprietary information, including our trade secrets, and we may not be able to obtain adequate remedies for such breaches. Enforcing a claim that a party illegally disclosed or misappropriated a trade secret is difficult, expensive and time-consuming, and the outcome is unpredictable. In addition, some courts inside and outside the United States are less willing or unwilling to protect trade secrets. We may also need to share our trade secrets and proprietary know-how with current or future partners, collaborators, contractors and others located in countries at heightened risk of theft of trade secrets, including through direct intrusion by private parties or foreign actors, and those affiliated with or controlled by state actors. As a result, we may encounter significant problems in protecting and defending our intellectual property both in the United States and abroad. In addition, competitors could purchase our therapeutics, if commercialized, and attempt to replicate or improve some or all of the competitive advantages we derive from our development efforts, reverse engineer our technologies, and design their products around our protected technologies or develop their own competitive technologies that fall outside of our intellectual property rights.
Moreover, third parties may still obtain this information or may come upon this or similar information independently, and we would have no right to prevent them from using that technology or information to compete with us. If any of these events occurs or if we otherwise lose protection for our trade secrets, the value of this information may be greatly reduced, and our competitive position could be harmed. If we do not apply for patent protection prior to such publication or if we cannot otherwise maintain the confidentiality of our proprietary technology and other confidential information, then our ability to obtain patent protection or to protect our trade secret information may be jeopardized.
We may become subject to claims challenging the inventorship or ownership of our patents and other intellectual property.
We may be subject to claims that former employees, collaborators or other third parties have an interest in our patents or other intellectual property as an inventor or co-inventor. The failure to name the proper inventors on a patent application can result in the patents issuing thereon being invalid or unenforceable. Inventorship disputes may arise from conflicting views regarding the contributions of different individuals named as inventors, the effects of foreign laws where foreign nationals are involved in the development of the subject matter of the patent, conflicting obligations of third parties involved in developing our current or future product candidates or as a result
48
Table of Contents
of questions regarding co-ownership of potential joint inventions. For example, we may have inventorship disputes arise from conflicting obligations of consultants or others who are involved in developing our product candidates. Litigation may be necessary to resolve these and other claims challenging inventorship or ownership. Alternatively, or additionally, we may enter into agreements to clarify the scope of our rights in such intellectual property. If we fail in defending any such claims, in addition to paying monetary damages, we may lose valuable intellectual property rights, such as exclusive ownership of, or right to use, valuable intellectual property. Such an outcome could have a material adverse effect on our business, financial condition, results of operations, and prospects. Even if we are successful in defending against such claims, litigation could result in substantial costs and be a distraction to management and other employees.
In addition, while it is our policy to require our employees and contractors who may be involved in the conception or development of intellectual property to execute agreements assigning such intellectual property to us, we may be unsuccessful in executing such an agreement with each party who, in fact, conceives or develops intellectual property that we regard as our own. The assignment of intellectual property rights may not be self-executing, or the assignment agreements may be breached, and we may be forced to bring claims against third parties, or defend claims that they may bring against us, to determine the ownership of what we regard as our intellectual property. Such claims could have a material adverse effect on our business, financial condition, results of operations and prospects.
Patent terms may be insufficient to protect our competitive position on any current and future product candidates for an adequate amount of time.
Patents have a limited lifespan. In the United States, if all maintenance fees are timely paid, the natural expiration of a patent is generally 20 years from its earliest U.S. non-provisional filing date. Various patent term adjustments or extensions may be available, but the life of a patent, and the protection it affords, is limited. Even if patents covering our current or future product candidates are obtained, once the patent life has expired, we may be open to competition from competitive products, including biosimilars or interchangeables. Given the amount of time required for the development, testing and regulatory review of new product candidates, patents protecting such candidates might expire before or shortly after such candidates are commercialized. As a result, our owned patent portfolio and any future licensed patent portfolio may not provide us with sufficient rights to exclude others from commercializing products similar or identical to ours.
If we do not obtain patent term extension for any product candidates we may develop, our business may be harmed.
Depending upon the timing, duration and specifics of any FDA regulatory approval of any of our current or future product candidates we may develop and our technology, our existing U.S. patents or one or more U.S. patents that may issue in the future based on a patent application that we own or may license may be eligible for limited patent term extension under the Hatch-Waxman Amendments. The Hatch-Waxman Amendments permit a patent extension term of up to five years as compensation for patent term lost during the FDA regulatory review process. A patent term extension cannot extend the remaining term of a patent beyond a total of 14 years from the date of product approval, only one patent may be extended and only those claims covering the approved product, a method for using it or a method for manufacturing it may be extended. The application for the extension must be submitted prior to the expiration of the patent for which extension is sought and within 60 days of FDA approval. A patent that covers multiple products for which approval is sought can only be extended in connection with one of the approvals. However, we may not be granted an extension because of, for example, failing to exercise due diligence during the testing phase or regulatory review process, failing to apply within applicable deadlines, failing to apply prior to expiration of relevant patents or otherwise failing to satisfy applicable requirements. Moreover, the applicable time period or the scope of patent protection afforded could be less than we request. In addition, to the extent we wish to pursue patent term extension based on a patent that we may in-license from a third party, we would need the cooperation of that third party. If we are unable to obtain patent term extension or the term of any such extension is less than we request, our competitors may obtain approval of competing products following our patent expiration, and our revenue could be reduced. Any of the foregoing could have a material adverse effect on our business, financial condition, results of operations, and prospects.
49
Table of Contents
If our trademarks and trade names are not adequately protected, then we may not be able to build name recognition in our markets of interest and our business may be adversely affected.
Our current or future registered or unregistered trademarks or trade names may be challenged, infringed, diluted, circumvented or declared generic or descriptive or determined to be infringing, misappropriating or violation on other marks. We may not be able to protect our rights to these trademarks and trade names or may be forced to stop using these names, which we need for name recognition by potential partners or customers in our markets of interest.
During trademark registration proceedings, we may receive rejections of our applications by the USPTO or in other foreign jurisdictions. Although we would be given an opportunity to respond to those rejections, we may be unable to overcome such rejections. In addition, in the USPTO and in comparable agencies in many foreign jurisdictions, third parties are given an opportunity to oppose pending trademark applications and to seek to cancel registered trademarks. Opposition or cancellation proceedings may be filed against our trademarks, and our trademarks may not survive such proceedings. If we are unable to establish name recognition based on our trademarks and trade names, we may not be able to compete effectively and our business may be adversely affected. We may license our trademarks and trade names to third parties, such as distributors. Although these license agreements may provide guidelines for how our trademarks and trade names may be used, a breach of these agreements or misuse of our trademarks and tradenames by our licensees may jeopardize our rights in or diminish the goodwill associated with our trademarks and trade names. Moreover, any name we have proposed to use with our current or future product candidates in the United States must be approved by the FDA, regardless of whether we have registered it, or applied to register it, as a trademark. Similar requirements exist in foreign jurisdictions. The FDA typically conducts a review of proposed product names, including an evaluation of potential for confusion with other product names. If the FDA (or an equivalent administrative body in a foreign jurisdiction) objects to any of our proposed proprietary product names, we may be required to expend significant additional resources in an effort to identify a suitable substitute name that would qualify under applicable trademark laws, not infringe, misappropriate or otherwise violate the existing rights of third parties and be acceptable to the FDA. Furthermore, in many countries, owning and maintaining a trademark registration may not provide an adequate defense against a subsequent infringement claim asserted by the owner of a senior trademark. We may not be able to obtain, protect or enforce our rights to these trademarks and trade names, which we need to build name recognition among potential partners or customers in our markets of interest. At times, competitors or other third parties may adopt trade names or trademarks similar to ours, thereby impeding our ability to build brand identity and possibly leading to market confusion. In addition, there could be potential trade name or trademark infringement, misappropriation, dilution or other claims brought by owners of other registered trademarks or trademarks that incorporate variations of our registered or unregistered trademarks or trade names. If we assert trademark infringement claims, a court may determine that the marks we have asserted are invalid or unenforceable, or that the party against whom we have asserted trademark infringement has superior rights to the marks in question. In this case, we could ultimately be forced to cease use of such trademarks.
Risks Related to Our Reliance on Third Parties
We are dependent on sole source and limited source suppliers for certain drug products, raw materials, samples, components, and other materials used in our product candidates. If we are unable to source these supplies on a timely basis, or establish longer-term contracts with our CMOs, we will not be able to complete our clinical trials on time and the development of our product candidates may be delayed.
We are dependent on sole source and limited source suppliers for certain drug products, raw materials, samples, components, and other materials used in our product candidates. We believe we would be able to source these supplies from alternative suppliers without materially and adversely impacting our business. If, however, we are unable to source these supplies on a timely basis, or establish longer-term contracts with our CMOs, we will not be able to complete our clinical trials on time and the development of our product candidates may be delayed. We do not currently have long-term supply contracts with any of our CMOs and they are not obligated to supply drug products to us for any period, in any specified quantity or at any certain price beyond the delivery contemplated by the relevant purchase orders. As a result, our suppliers could stop selling to us at commercially reasonable prices, or at all. While we intend to enter into long-term master supply agreements with certain of our CMOs in the future as we advance our clinical trials or commercialization plans, we may not be successful in negotiating such agreements
50
Table of Contents
on favorable terms or at all. If we do enter into such long-term master supply agreements, or enter into such agreements on less favorable terms than we currently have with such manufacturers, we could be subject to binding long-term purchase obligations that may be harmful to our business, including in the event that we do not conduct our trials on planned timelines or utilize the drug products that we are required to purchase. Any change in our relationships with our CMOs or changes to contractual terms of our agreements with them could adversely affect our business, financial condition, results of operations, and prospects. Furthermore, any of the sole source and limited source suppliers upon whom we rely could stop producing our supplies, cease operations or be acquired by, or enter into exclusive arrangements with, one or more of our competitors.
The drug substance (DS) and drug product (DP) for our product candidates are manufactured via conventional biopharmaceutical processing procedures, employing commonly-used and commercially available excipients and packaging materials. The DS and DP are required to be manufactured in compliance with multi-jurisdictional regulatory standards including current good manufacturing practices (cGMP) and comparable foreign requirements. Additionally, our manufacturing processes for ipsoprubart and ELA822 require special equipment, and identifying additional suppliers able to fabricate such equipment at their facility at acceptable costs may be difficult. Establishing additional or replacement suppliers for these supplies, and obtaining regulatory clearance or approvals that may result from adding or replacing suppliers, could take a substantial amount of time, result in increased costs and impair our ability to produce our products, which would adversely impact our business, financial condition, results of operations, and prospects. Any such interruption or delay may force us to seek similar supplies from alternative sources, which may not be available at reasonable prices, or at all. Any interruption in the supply of sole source or limited source components for our product candidates would adversely affect our ability to meet scheduled timelines and budget for the development and commercialization of our product candidates, could result in higher expenses and would harm our business. For example, following an FDA inspection ended in March 2026, the CMO facility we utilize for drug product manufacture received an FDA Form-483 detailing several adverse manufacturing and product quality observations. The FDA ultimately classified this inspection as “Official Action Indicated” (OAI), meaning the FDA deemed this site to be in unacceptable state of compliance. We understand this facility is undergoing a remediation program that will require a facility shutdown. While we understand that the Form-483 observations were not specific to our product candidates and we have not identified any product quality concerns specific to our material, the facility will be required to undergo remedial efforts in order to lift the OAI classification, including a temporary facility shutdown, and the timeline for our CMO to resolve these inspectional issues is still uncertain. Furthermore, unless the FDA removes the OAI status from this facility, the FDA may delay approval of any BLA that includes this facility as a product manufacturing site, which could delay regulatory approval of ipsoprubart. Although we have not yet experienced any significant disruption as a result of our reliance on limited or sole source suppliers, we have a limited operating history and cannot assure you that we will not experience disruptions in our supply chain in the future as a result of such reliance or otherwise.
We rely, and intend to continue to rely, on third parties to conduct our clinical trials and perform some of our research and potential nonclinical studies. If these third parties do not satisfactorily carry out their contractual duties, fail to comply with applicable regulatory requirements or do not meet expected deadlines, our development programs may be delayed or subject to increased costs or we may be unable to obtain marketing authorization, each of which may have an adverse effect on our business, financial condition, results of operations, and prospects.
We do not have the ability to independently conduct all aspects of our clinical trials ourselves. As a result, we are dependent on third parties to conduct our ongoing and planned clinical trials of ipsoprubart, ELA822, any nonclinical studies and all clinical trials of any future product candidates. The timing of the initiation and completion of these trials will therefore be partially controlled by such third parties and may result in delays to our development programs. Specifically, we expect CROs, independent clinical investigators and consultants to play a significant role in the conduct of these trials and the subsequent collection and analysis of data. However, these investigators, CROs and other third parties are not our employees, and we will not be able to control all aspects of their activities. Nevertheless, we are responsible for ensuring that each clinical trial is conducted in accordance with the applicable protocol and legal, regulatory and scientific standards, and our reliance on the investigators, CROs and other third parties does not relieve us of our regulatory responsibilities. We and our CROs are required to comply with GCP requirements, which are regulations and guidelines enforced by the FDA and comparable foreign regulatory authorities for product candidates in clinical development. Regulatory authorities enforce these GCP requirements
51
Table of Contents
through periodic inspections of trial sponsors, clinical trial investigators and clinical trial sites. If we or any of our CROs or clinical trial sites fail to comply with applicable GCP requirements, the data generated in our clinical trials may be deemed unreliable, and the FDA or comparable foreign regulatory authorities may require us to perform additional clinical trials before approving our marketing applications. We cannot assure you that, upon inspection, the FDA or comparable foreign regulatory authorities will determine that our clinical trials comply with GCPs. In addition, our clinical trials must be conducted with materials produced under cGMP regulations. Our failure or the failure of third parties on whom we rely to comply with these regulations may require us to stop and/or repeat clinical trials, which would delay the marketing authorization process.
There is no guarantee that any such CROs, clinical trial investigators or other third parties on which we rely will devote adequate time and resources to our development activities or perform as contractually required. In addition, these third parties may be subject to supply chain or inflationary pressures that limit their ability to achieve anticipated timelines or result in a greater cost to us. For example, we are aware of recurrent shortages of NHPs available for nonclinical studies and although that is not expected to impact our current business, if we begin new product development programs we could be subject to longer development times or difficulty completing necessary research. If any of these third parties fail to meet expected deadlines, adhere to our clinical protocols or meet regulatory requirements, otherwise perform in a substandard manner, or terminate their engagements with us, the timelines for our development programs may be extended or delayed or our development activities may be suspended or terminated. If our clinical trial site terminates for any reason, we may experience the loss of follow-up information on participants enrolled in such clinical trial unless we are able to transfer those participants to another qualified clinical trial site, which may be difficult or impossible.
In addition, our CROs have the right to terminate their agreements with us in the event of an uncured material breach and under other specified circumstances. If any of our relationships with these third parties terminate, we may not be able to enter into arrangements with alternative third parties on commercially reasonable terms or at all. Switching or adding additional CROs, investigators and other third parties involves additional cost and requires our management’s time and focus. In addition, there is a natural transition period when a new CRO commences work. As a result, delays occur, which can materially impact our ability to meet our desired clinical development timelines.
Furthermore, these third parties may also have relationships with other entities, some of which may be our competitors for whom they may also be conducting clinical trials or other product candidate development activities that could harm our competitive position. If these third parties do not successfully carry out their contractual duties, meet expected deadlines or conduct our clinical trials in accordance with regulatory requirements or our stated protocols, we will not be able to obtain, or may be delayed in obtaining, regulatory approval for ipsoprubart, ELA822 and any future product candidates and will not be able to, or may be delayed in our efforts to, successfully commercialize our products.
We rely on third parties to manufacture our product candidates and clinical product supplies and we may not be able to obtain adequate supplies at a reasonable cost or in a timely way.
The process of manufacturing product candidates is complex and highly regulated. We do not have any manufacturing facilities or plans to establish manufacturing capabilities in the near future. We rely, and expect to continue to rely, on third parties, including foreign manufacturers, for the manufacture of our product candidates for nonclinical and clinical testing, development purposes, to support marketing application submissions, as well as for commercial manufacture if any of our product candidates obtain regulatory approval. This reliance on third parties increases the risk that we will not have sufficient quantities of our product candidates or such quantities at an acceptable cost or quality, which could delay, prevent or impair our development or commercialization efforts. In addition, global health crises or geopolitical conflict may result in disruptions to the operations or an extended shutdown of certain businesses, which could include certain of our contract manufacturers. Further, as our product candidates are developed through nonclinical studies to late-stage clinical trials towards approval and commercialization, we are likely to alter various aspects of the development program, such as manufacturing and testing methods, to optimize processes and results, which may result in additional cost or delay.
We have only limited supply arrangements in place with respect to our product candidates, and these arrangements do not extend to commercial supply. We acquire many key materials on a purchase order basis. As a result, we may
52
Table of Contents
not have long-term committed arrangements with respect to aspects of our product candidates and other materials. We will need to establish one or more agreements with third parties to develop and scale up the drug manufacturing process, conduct testing, and generate data to support regulatory submissions, and may be unable to do so on favorable terms. If we obtain regulatory approval for any of our product candidates, we will need to establish an agreement for commercial manufacture with a third party. Even if we are able to establish agreements with third-party manufacturers, reliance on third-party manufacturers entails additional risks, including, but not limited to:
| ∎ | reliance on the third party for regulatory, compliance and quality assurance; |
| ∎ | reliance on the third party for product development, analytical testing, and data generation to support regulatory applications; |
| ∎ | operations of our third-party manufacturers or suppliers could be disrupted by conditions unrelated to our business or operations, including the bankruptcy of the manufacturer or supplier, the issuance of an FDA Form 483 notice or warning letter, or other enforcement action by FDA or other comparable foreign regulatory authority; |
| ∎ | the possible breach of the manufacturing agreement by the third party; |
| ∎ | the possible infringement, misappropriation, violation or unauthorized disclosure of our intellectual property and proprietary rights, including our trade secrets, know-how and other confidential information; |
| ∎ | the possible termination or nonrenewal of the agreement by the third party at a time that is costly or inconvenient for us; |
| ∎ | competition with other companies for access to manufacturing capacity; |
| ∎ | carrier disruptions or increased costs that are beyond our control; and |
| ∎ | failure to deliver our products under specified storage conditions and in a timely manner. |
Third-party manufacturers may not be able to comply with cGMP regulations or similar regulatory requirements outside of the United States. Our CMOs, any potential future collaborators and their CMOs could be subject to periodic unannounced inspections by the FDA and other comparable foreign regulatory authorities, to monitor and ensure compliance with cGMP. Despite our efforts to audit and verify regulatory compliance, we have limited control over the ability of third-party manufacturers to maintain adequate quality control, quality assurance and qualified personnel, and one or more of our third-party manufacturing vendors may be found on regulatory inspection by the FDA or other comparable foreign regulatory authorities to be noncompliant with cGMP and comparable foreign regulations. For example, following an inspection ended in March 2026, the CMO facility we utilize for drug product manufacture received an FDA Form-483, ultimately resulting in an OAI classification. While we understand that the Form-483 observations were not specific to our product candidates, the facility will be required to undergo significant remedial efforts in order to lift the OAI classification, including a temporary facility shutdown, and there is no guarantee that our CMO will be able to resolve these inspectional issues in a timely manner.
If the FDA determines that our CMOs are not in compliance with FDA laws and regulations, including those governing cGMPs, the FDA may not approve a BLA until the deficiencies are corrected or we replace the manufacturer in our application with a manufacturer that is in compliance. Comparable risks apply abroad. Moreover, our failure, or the failure of our third-party manufacturers and suppliers, to comply with applicable regulations could result in sanctions being imposed on us, including clinical holds, fines, injunctions, civil penalties, seizures or recalls of product candidates, operating restrictions and criminal prosecutions, any of which could significantly and adversely affect supplies of our products. In addition, approved products and the facilities at which they are manufactured are required to maintain ongoing compliance with extensive FDA requirements and comparable foreign requirements, including ensuring that quality control and manufacturing procedures conform to cGMP requirements. As such, our CMOs are subject to continual review and periodic inspections to assess compliance with cGMPs. Furthermore, although we do not have day-to-day control over the operations of our CMOs, we are responsible for ensuring compliance with applicable laws and regulations, including cGMPs.
Further, we rely on third parties located in China for some of our contract manufacturing, and we expect to continue to use such third-party manufacturers for such purposes. For any activities conducted in China, we are exposed to the possibility of product supply disruption and increased costs in the event of changes in the policies of the United States or Chinese governments, political unrest or unstable economic conditions in China. In addition, certain
53
Table of Contents
Chinese biotechnology companies may become subject to trade restrictions, sanctions, other regulatory requirements, or proposed legislation by the U.S. government, which could restrict or even prohibit our ability to work with such entities, thereby potentially disrupting the supply of material to us. For example, the BIOSECURE Act, which was recently signed into law in December 2025 as part of the National Defense Authorization Act for FY 2026, prohibits U.S. federal agencies from procuring or obtaining biotechnology equipment or services produced or provided by a “biotechnology company of concern” (BCC); entering into, extending, or renewing government contracts with an entity that directly or indirectly (e.g., via a subcontractor) uses biotechnology equipment or services from a BCC in performance of that federal contract; and/or issuing grants or loans to purchase, obtain, or use biotechnology equipment or services produced by a BCC. The BIOSECURE Act also prohibits U.S. government loan and grant recipients from using federal loan or grant money to enter into contracts with entities that use equipment from BCCs in the performance of any federal prime contract or subcontract. Companies designated as a BCC include those that are identified on the U.S. Department of Defense’s annual List of Chinese Military Companies, also known as the 1260H List, and the U.S. Government also has the ability to designate entities as BCCs through a separate designation process. There is a safe harbor provision providing that the restrictions do not apply to equipment or services that were formerly but are no longer provided by a “biotechnology company of concern,” as well as a grandfathering provision providing that the prohibitions shall not apply for a five-year period to biotechnology equipment or services produced or provided under a contract or agreement entered into before the applicable effective date; the BIOSECURE Act also has carveouts for manufacture of drugs for supply under Medicaid and Medicare Part B, subject to the Secretary of Veteran Affairs’ discretion. It is unclear whether the grandfathering provision and carveouts would apply to entities designated as “biotechnology companies of concern” due to their inclusion on the 1260H List. The guidance to be issued by the Office of Management and Budget within the next year regarding implementation of the BIOSECURE Act may provide further clarity on this point. We are currently a party to agreements with WuXi Biologics (WuXi), pursuant to which WuXi provides clinical development and manufacturing services to us related to ELA822. WuXi may become designated as a “biotechnology company of concern” through periodic updates of the 1260H List, or may also be designated as such by the U.S. Government through the process outlined in the BIOSECURE Act. If this law, or similar laws that are passed impact WuXi or other Chinese biotechnology manufacturing companies that may become contractors of ours or provide biotechnology equipment or services in the manufacture of our products or product candidates, we may be restricted in our ability to work with WuXi and other Chinese biotechnology manufacturing companies to the extent we would contract with, or otherwise receive funding from, the U.S. government. As a result, we may need to seek alternative CMO relationships. While we believe we will be able to identify and contract with such alternative CMOs, we cannot predict the terms of any such alternative arrangement nor what actions may ultimately be taken with respect to trade relations between the United States and China or other countries, what products and services may be subject to such actions or what actions may be taken by China or the other countries in retaliation. In addition, any unfavorable government policies on international trade, such as export controls, capital controls or tariffs, new legislation or regulations, renegotiation of existing trade agreements, or any retaliatory trade actions due to recent or future trade tension, may impede, delay, limit, or increase the cost of manufacturing our product candidates. Such events could result in our clinical or commercial supply of drug, packaging and other services being interrupted or limited, which could harm our business.
In addition, our third-party manufacturers and suppliers are subject to numerous environmental, health and safety laws and regulations, including those governing the handling, use, storage, treatment and disposal of waste products, and failure to comply with such laws and regulations could result in significant costs associated with civil or criminal fines and penalties for such third parties. Based on the severity of regulatory actions that may be brought against these third parties in the future, our clinical or commercial supply of drug and packaging and other services could be interrupted or limited, which could harm our business.
Any performance failure on the part of our existing or future manufacturers could delay clinical development or regulatory approval. We do not currently have arrangements in place for redundant supply or a second source for bulk DS. If our current CMOs for preclinical and clinical testing cannot perform as agreed, we may be required to replace such CMOs. Although we believe that there are several potential alternative manufacturers who could manufacture our product candidates, we may incur added costs and delays in identifying and qualifying any such replacement manufacturer or be able to reach agreement with any alternative manufacturer. Further, our third-party manufacturers may experience manufacturing or shipping difficulties due to resource constraints or as a result of
54
Table of Contents
| natural disasters, labor disputes, unstable political environments, or global health crises. If our current third-party manufacturers cannot perform as agreed, we may be required to replace such manufacturers and we may be unable to replace them on a timely basis or at all. |
Our current and anticipated future dependence upon others for the manufacture of our product candidates may adversely affect our future profit margins and our ability to commercialize any products that obtain regulatory approval on a timely and competitive basis.
We may, in the future, seek to enter into collaborations with third parties for the discovery, development and commercialization of product candidates, if approved, and we may not be successful in doing so. If those collaborations are not successful, we may not be able to capitalize on the market potential of ipsoprubart, ELA822 or future product candidates.
We may seek third-party collaborators for the development and commercialization of our current or any future product candidates, if approved, on a select basis, including potentially in specific foreign jurisdictions. We have not entered into any such collaborations to date. Our likely collaborators for any future collaboration arrangements include large and mid-size biopharmaceutical companies, regional and national biopharmaceutical companies and biotechnology companies. We will face significant competition in seeking appropriate collaborators. Whether we decide to enter into any collaboration arrangement will depend, among other things, on our assessment of the future collaborator’s resources and expertise, the terms and conditions of the proposed collaboration and the proposed collaborator’s evaluation of our business. As such, there can be no assurance that we will reach a definitive agreement for a future collaboration with any third-party collaborators.
If we do enter into any such arrangements with any third parties, we will likely have limited control over the amount and timing of resources that our future collaborators dedicate to the development or commercialization of our current or any future product candidates. Our ability to generate revenues from these arrangements will be dependent on our future collaborators’ abilities and efforts to successfully perform the functions assigned to them in these arrangements. Collaborations with future collaborators involving our current or any future product candidates would pose numerous risks to us, including the following:
| ∎ | collaborators have significant discretion in determining the efforts and resources that they will apply to these collaborations and may not perform their obligations as expected; |
| ∎ | collaborators may de-emphasize or not pursue development and commercialization of our current or any future product candidates or may elect not to continue or renew development or commercialization programs based on clinical trial results, changes in the collaborators’ strategic focus, including as a result of a sale or disposition of a business unit or development function, or available funding or external factors such as an acquisition that diverts resources or creates competing priorities; |
| ∎ | collaborators may delay clinical trials, provide insufficient funding for a clinical trial program, stop a clinical trial or abandon a product candidate, repeat or conduct new clinical trials or require a new formulation of a product candidate for clinical testing; |
| ∎ | collaborators could independently develop, or develop with third parties, products that compete directly or indirectly with our current or any future product candidates if the collaborators believe that competitive products are more likely to be successfully developed or can be commercialized under terms that are more economically attractive than ours; |
| ∎ | a collaborator with marketing and distribution rights to multiple products may not commit sufficient resources to the marketing and distribution of our product, if approved, relative to other products; |
| ∎ | collaborators may not properly obtain, maintain, defend or enforce our intellectual property rights or may use our proprietary information and intellectual property in such a way as to invite litigation or other intellectual property related proceedings that could jeopardize or invalidate our proprietary information and intellectual property or expose us to potential litigation or other intellectual property related proceedings; |
| ∎ | disputes may arise between the collaborators and us, such as conflicts concerning the interpretation of preclinical or clinical data, the achievement of milestones, the interpretation of contractual obligations, payments for services, development obligations or the ownership of intellectual property developed during our collaboration, any of which could result in the delay or termination of the research, development or, if |
55
Table of Contents
| approved, commercialization of our current or any future product candidates or that result in costly litigation or arbitration that diverts management attention and resources and in turn could prevent us from generating revenue and limit or prevent us from entering into additional collaborations; |
| ∎ | collaborations may be terminated and, if terminated, may result in a need for additional capital to pursue further development or, if approved, commercialization of the applicable product candidates; |
| ∎ | collaboration agreements may not lead to development or, if approved, commercialization of product candidates in the most efficient manner or at all; and |
| ∎ | if a future collaborator of ours were to be involved in a business combination, the continued pursuit and emphasis on our product development or, if approved, commercialization program could be delayed, diminished or terminated. |
If we establish one or more collaborations, all of the risks relating to product development, regulatory approval and, if approved, commercialization described above would also apply to the activities of any such future collaborators.
Risks Related to Government Regulation
The regulatory approval process is highly uncertain, and we may be unable to obtain, or may be delayed in obtaining, U.S. or foreign regulatory approval and, as a result, unable to commercialize ipsoprubart, ELA822 and any future product candidates. Even if we believe our current, or planned clinical trials are successful, regulatory authorities may not agree that they provide adequate data on safety or efficacy.
Ipsoprubart, ELA822 and any future product candidates are subject to extensive governmental regulations relating to, among other things, research, testing, development, manufacturing, approval, recordkeeping, reporting, labeling, storage, packaging, advertising and promotion, pricing, post-approval monitoring, marketing and distribution of products. Rigorous nonclinical testing and clinical trials and an extensive regulatory approval process are required to be completed successfully in the United States and in many foreign jurisdictions before a new biopharmaceutical product can be marketed. Satisfaction of these and other regulatory requirements is costly, time consuming, uncertain and subject to unanticipated delays. It is possible that none of the product candidates we may develop will obtain the regulatory approvals necessary for us to begin selling them.
In the U.S., we are not permitted to market our product candidates in the U.S. until we receive regulatory approval of a BLA from the FDA. Prior to obtaining approval to commercialize a product candidate in the U.S. or abroad, we must demonstrate with substantial evidence from adequate and well-controlled clinical trials, and to the satisfaction of the FDA or comparable foreign regulatory authorities, that such product candidates are safe and effective for their intended uses, and in the case of biological products in the U.S., that such product candidates are safe, pure, and potent for their intended uses. The time required to obtain FDA and other approvals is unpredictable but typically takes many years following the commencement of clinical trials, depending on the type, complexity and novelty of the product candidate. The standards that the FDA and its foreign counterparts use when regulating us require judgment and can change, which makes it difficult to predict with certainty their application. Any analysis we perform of data from nonclinical and clinical activities is subject to confirmation and interpretation by regulatory authorities, which could delay, limit or prevent regulatory approval. Even if we believe available nonclinical or clinical data support the safety purity, potency or efficacy of our product candidates, such data may not be sufficient to obtain approval from the FDA and comparable foreign regulatory authorities. The FDA or comparable foreign regulatory authorities, as the case may be, may also require us to conduct additional nonclinical studies or clinical trials for our product candidates either prior to or post-approval, or may object to elements of our clinical development program.
The FDA or comparable foreign regulatory authorities can delay, limit or deny approval of a product candidate for many reasons, including:
| ∎ | such authorities may disagree with the design or execution of our clinical trials; |
| ∎ | negative or ambiguous results from our clinical trials or results may not meet the level of statistical significance or persuasiveness required by the FDA or comparable foreign regulatory agencies for approval; |
| ∎ | serious and unexpected drug-related side effects may be experienced by patients in our clinical trials or by individuals using drugs similar to our product candidates; |
56
Table of Contents
| ∎ | the population studied in the clinical trial may not be sufficiently broad or representative to assure safety in the full population for which we seek approval; |
| ∎ | such authorities may not accept clinical data from trials that are conducted at clinical facilities or in countries where the standard of care (SOC) is potentially different from that of their own country; |
| ∎ | we may be unable to demonstrate that a product candidate’s clinical and other benefits outweigh its safety risks; |
| ∎ | such authorities may disagree with our interpretation of data from nonclinical studies or clinical trials; |
| ∎ | such authorities may not agree that the data collected from clinical trials of our product candidates are acceptable or sufficient to support the submission of a BLA, or other submission or to obtain regulatory approval in the U.S. or elsewhere, and such authorities may impose requirements for additional nonclinical studies or clinical trials; |
| ∎ | such authorities may disagree with us regarding the formulation, labeling and/or the product specifications of our product candidates; |
| ∎ | approval may be granted only for indications that are significantly more limited than those sought by us, and/or may include significant restrictions on distribution and use; |
| ∎ | such authorities may find deficiencies in the manufacturing processes or facilities of the third-party manufacturers with which we contract for clinical and commercial supplies; or |
| ∎ | such authorities may not accept a submission due to, among other reasons, the content or formatting of the submission. |
In addition, the approval policies or regulations of the FDA and other comparable regulatory authorities in other jurisdictions may change in a manner rendering our clinical data insufficient for approval. Any delay or failure in obtaining required approvals could have a material and adverse effect on our ability to generate revenues from the particular product candidate for which we are seeking approval. Furthermore, any regulatory approval to market a product may be subject to limitations on the approved uses for which we may market the product or on the labeling or other restrictions.
We are also subject to or may in the future become subject to numerous foreign regulatory requirements governing, among other things, the conduct of clinical trials, manufacturing and marketing authorization, pricing and third-party reimbursement. The foreign regulatory approval process varies among countries and may include all of the risks associated with the FDA approval process described above, as well as risks attributable to the satisfaction of local regulations in foreign jurisdictions. In the European Union (EU) and the United Kingdom, approval pathways, procedural timetables and post-authorization requirements may differ from those in the United States and may require separate national, centralized or other jurisdiction-specific processes, which may increase uncertainty, cost and delay.
Moreover, the time required to obtain approval may differ from that required to obtain FDA approval. Obtaining regulatory approval outside of the United States, including in the EU, is a lengthy and expensive process. To obtain marketing authorization from the European Commission through the centralized procedure, following the opinion of the EMA, for any product candidate, we would need to submit a Marketing Authorization Application (MAA) and satisfy extensive requirements regarding safety, efficacy, and quality. Even if a product candidate receives a marketing authorization, the European Commission or EMA may limit the indications for which the product may be marketed, require extensive warnings on the product labeling, or require expensive and time-consuming post-authorization studies or reporting as conditions of approval. Failure to obtain or maintain marketing authorization in the EU could limit our commercial opportunity and materially harm our business. FDA approval does not ensure approval by regulatory authorities outside the United States and vice versa. Any delay or failure to obtain United States or foreign regulatory approval for a product candidate could have a material and adverse effect on our business, financial condition, results of operations, and prospects.
We may not be able to obtain or maintain orphan drug designations for any of our product candidates, and we may be unable to maintain the benefits associated with orphan drug designation, including the potential for market exclusivity.
Regulatory authorities in some jurisdictions, including the United States, may designate drugs or biologics for relatively small patient populations as orphan drugs. Under the Orphan Drug Act, as amended, the FDA may designate a drug or
57
Table of Contents
biologic as an orphan product if it is intended to treat a rare disease or condition, which is generally defined as a patient population of fewer than 200,000 individuals in the United States, or a patient population of greater than 200,000 individuals in the United States, but for which there is no reasonable expectation that the cost of developing the drug will be recovered from sales in the United States. In the United States, orphan designation entitles a party to financial incentives such as opportunities for grant funding toward clinical trial costs, tax advantages and user-fee waivers. In addition, if a product candidate that has orphan designation subsequently receives the first FDA approval for the disease or condition for which it has such designation, the product is entitled to orphan drug exclusivity, which means that the FDA may not approve any other applications, including an NDA or BLA, to market the same product for the same approved indication or use within the relevant rare disease or condition for seven years, except in limited circumstances, such as a showing of clinical superiority to the product with orphan drug exclusivity within the relevant indication or use or where the manufacturer is unable to assure sufficient product quantity to meet the needs relating to the approved indication or use of patients with the orphan disease or condition.
In October 2024, the FDA granted orphan drug designation to ipsoprubart for the treatment of patients with HLH. If ipsoprubart receives regulatory approval for an indication broader than HLH, ipsoprubart may no longer be eligible for orphan exclusivity. In addition, we may seek orphan drug designations for our other product candidates, but there can be no assurances that we will be able to obtain such designations. Even if we obtain orphan drug designation for a product candidate, we may not be able to obtain orphan drug exclusivity for that product candidate, if approved. Further, even if we obtain orphan drug exclusivity for a product, that exclusivity may not effectively protect the product from competition because different drugs can be approved for the same indications and uses. Even after an orphan drug is approved, the FDA can subsequently approve the “same drug,” as defined by FDA, for the same indication or use if the FDA concludes that the later drug is clinically superior because it is shown to be safer, more effective or makes a major contribution to patient care within the exclusivity-protected indication. Orphan drug exclusivity in the United States may also be lost if the FDA later determines that the initial request for designation was materially defective. In addition, orphan drug exclusivity does not prevent the FDA from approving competing drugs containing different active ingredients for the same or similar indication. If a subsequent drug is approved for marketing for the same or a similar indication or use as any of our product candidates that receive regulatory approval, we may face increased competition and lose market share regardless of orphan drug exclusivity. Orphan drug designation neither shortens the development time nor regulatory review time of a drug or biologic nor gives the drug or biologic any advantage in the regulatory review or approval process.
A Fast Track Designation from the FDA, even if granted for any of our product candidates, may not lead to a faster development or regulatory review or approval process, and does not increase the likelihood that our product candidates will receive regulatory approval.
In October 2024, the FDA granted Fast Track designation for ipsoprubart for the treatment of sHLH. Depending on the data from our nonclinical studies and clinical trials, we may decide to seek such designation for some or all of our other product candidates. The Fast Track program is intended to expedite or facilitate the process for reviewing product candidates that meet certain criteria. Specifically, drugs and biologics are eligible for Fast Track designation if they are intended, alone or in combination with one or more drugs or biologics, to treat a serious or life-threatening disease or condition and demonstrate the potential to address unmet medical needs for the disease or condition. Fast Track designation applies to the combination of the product candidate and the specific indication for which it is being studied. The sponsor of a Fast Track product candidate has opportunities for more frequent interactions with the applicable FDA review team during product development and, once a BLA is submitted, the application may be eligible for priority review. A BLA submitted for a Fast Track product candidate may also be eligible for rolling review, where the FDA may consider for review sections of the BLA on a rolling basis before the complete application is submitted, if the sponsor provides a schedule for the submission of the sections of the BLA, the FDA agrees to accept sections of the BLA and determines that the schedule is acceptable, and the sponsor pays any required user fees upon submission of the first section of the application.
The FDA has broad discretion whether or not to grant this designation. Even if we believe a particular product candidate is eligible for this designation, we cannot assure you that the FDA would decide to grant it. Even if we do receive Fast Track Designation for any of our product candidates, such product candidates may not experience a faster development process, review or approval compared to conventional FDA procedures. The FDA may also withdraw Fast Track Designation if it believes that the designation is no longer supported by data from our clinical development program. Furthermore, such a designation does not increase the likelihood that ipsoprubart or any other
58
Table of Contents
product candidate that may be granted Fast Track designation will receive regulatory approval in the U.S. Many product candidates that have received Fast Track Designation have ultimately failed to obtain approval.
A Breakthrough Therapy Designation by the FDA or PRIME designation by the EMA, even if granted for any of our product candidates, may not lead to a faster development or regulatory review or approval process, and does not increase the likelihood that our product candidates will receive regulatory approval.
The FDA has granted Breakthrough Therapy Designation for ipsoprubart for the treatment of sHLH, and the EMA has granted PRIME designation for ipsoprubart for the treatment of sHLH, and, in the future, we may seek Breakthrough Therapy Designation or PRIME designation for our other product candidates. A Breakthrough Therapy is defined as a drug or biologic that is intended, alone or in combination with one or more other drugs or biologics, to treat a serious or life-threatening disease or condition, and preliminary clinical evidence indicates that the drug or biologic may demonstrate substantial improvement over existing therapies with respect to one or more clinically significant endpoints, such as substantial treatment effects observed early in clinical development. For product candidates that have been designated as Breakthrough Therapies, interaction and communication between the FDA and the sponsor can help to identify the most efficient path for development. Breakthrough Therapy designation comes with the benefits of Fast Track designation, which means that the sponsor may submit sections of the NDA or BLA for review on a rolling basis, provided certain conditions are met. The FDA has broad discretion whether or not to grant this designation.
PRIME is a program launched by the EMA that provides early, proactive, and enhanced scientific and regulatory support for medicines under development that target an unmet medical need and are expected to be of major public health interest. This regulatory program offers enhanced interaction and early dialogue with the EMA and is designed to optimize development plans and speed evaluation ensuring these medicines reach patients as early as possible. The benefits of a PRIME designation include the appointment of a rapporteur before submission of a marketing authorization application, early dialogue and scientific advice at key development milestones, and the potential to qualify products for accelerated review earlier in the application process. The EMA has broad discretion whether or not to grant this designation.
Even if we believe a particular product candidate is eligible for these designations, we cannot assure you that the FDA or EMA would decide to grant them. Breakthrough Therapy and PRIME designations may not result in a faster development process, review or approval compared to conventional FDA or EMA procedures, respectively. In addition, the FDA may withdraw Breakthrough Therapy Designation if it believes that the designation is no longer supported by data from our clinical development program. The Breakthrough Therapy and PRIME designations do not assure ultimate regulatory approval by the FDA or the European Commission. Many drugs and biologics that have received Breakthrough Therapy or PRIME designation have failed to obtain approval.
We may be unable to maintain the benefits associated with our EU orphan designation, including orphan market exclusivity, which could limit the commercial prospects of our product candidates.
We have received orphan designation in the EU for one or more of our product candidates. Orphan designation entitles an applicant to incentives such as fee reductions or fee waivers, protocol assistance, and access to the centralized marketing authorization procedure. Upon grant of a marketing authorization, orphan medicinal products are entitled to a ten-year period of market exclusivity for the approved therapeutic indication, during which competent authorities cannot accept another marketing authorization application or accept an application to extend for a similar product or cannot grant a marketing authorization for the same indication. However, orphan market exclusivity may be reduced to six years if, at the end of the fifth year, it is established that our product no longer meets the orphan designation criteria, including where it is determined that the product is sufficiently profitable not to justify maintenance of market exclusivity, or where the prevalence of the condition has increased above the threshold. Additionally, a marketing authorization may be granted to a similar product for the same indication during the ten-year period if another applicant can establish that its product is safer, more effective or otherwise clinically superior to ours. We cannot guarantee that we will maintain our orphan designation or benefit from the full period of market exclusivity, and any loss or reduction of such exclusivity could materially and adversely affect our commercial prospects in the EU.
Obtaining and maintaining regulatory approval of our product candidates in one jurisdiction does not mean that we will be successful in obtaining regulatory approval of our product candidates in other jurisdictions.
Obtaining and maintaining regulatory approval of our product candidates in one jurisdiction does not guarantee that we will be able to obtain or maintain regulatory approval in any other jurisdiction. For example, even if the FDA or
59
Table of Contents
European Commission grants regulatory approval of a product candidate, comparable regulatory authorities in foreign jurisdictions must also approve the manufacturing, marketing and promotion of the product candidate in those countries. However, a failure or delay in obtaining regulatory approval in one jurisdiction may have a negative effect on the regulatory approval process in others. Approval procedures vary among jurisdictions and can involve requirements and administrative review periods different from those in the United States, including additional nonclinical studies or clinical trials as clinical trials conducted in one jurisdiction may not be accepted by regulatory authorities in other jurisdictions. In many jurisdictions outside the United States, pricing and reimbursement approvals or decisions are separate from marketing authorization and may be required before a product can achieve broad commercial uptake in that jurisdiction. In some cases, the price that we intend to charge for our products is also subject to approval. Obtaining foreign regulatory approvals and compliance with foreign regulatory requirements could result in significant delays, difficulties and costs for us and could delay or prevent the introduction of our products in certain countries. If we or any partner we work with fail to comply with the regulatory requirements in international markets or fail to receive applicable regulatory approvals, our target market will be reduced and our ability to realize the full market potential of our product candidates will be harmed.
Even if we receive regulatory approval for ipsoprubart, ELA822 and any future product candidates, we will be subject to ongoing regulatory obligations and continued regulatory review, which may result in significant additional expense. Additionally, ipsoprubart, ELA822 and any future product candidates, if approved, could be subject to labeling and other restrictions and market withdrawal. We may also be subject to penalties if we fail to comply with regulatory requirements or experience unanticipated problems with our products.
Any regulatory approvals that we obtain for ipsoprubart, ELA822 or any of our future product candidates may also be subject to limitations on the approved indicated uses for which a product may be marketed or to the conditions of approval, or contain requirements for potentially costly post-marketing testing and surveillance to monitor the safety and efficacy of the product candidate. In addition, if the FDA or other comparable foreign regulatory authorities approve any of our future product candidates, the manufacturing processes, labeling, packaging, distribution, post-approval monitoring, pharmacovigilance obligations and adverse event reporting, storage, import, export, advertising, promotion and recordkeeping for the product will be subject to extensive and ongoing regulatory requirements. The FDA has significant post-market authority, including the authority to require labeling changes based on new safety information and to require post-market studies or clinical trials to evaluate safety risks related to the use of a product or to require withdrawal of the product from the market. The FDA also has the authority to require a REMS after approval, which may impose further requirements or restrictions on the distribution or use of an approved drug. The manufacturing facilities we use to make future product candidates, if any, will also be subject to periodic review and inspection by the FDA or other comparable foreign regulatory authorities, including for continued compliance with cGMP and comparable foreign requirements. The discovery of any new or previously unknown problems with our CMOs, manufacturing processes or facilities may result in restrictions on the product, manufacturer or facility, including withdrawal of the product from the market. If we rely on CMOs, we will not have control over compliance with applicable rules and regulations by such manufacturers.
If we or our manufacturers or service providers fail to comply with applicable continuing regulatory requirements in the United States or foreign jurisdictions in which we seek to market our products, we or they may be subject to, among other things:
| ∎ | Form 483s, restrictions on the manufacturing of the product, product recalls or withdrawal of the product from the market; |
| ∎ | warning or untitled letters or holds on clinical trials; |
| ∎ | refusal of the FDA or comparable foreign regulatory authorities to accept new marketing applications or approve pending applications or supplements to approved applications, or suspension, variation or revocation of product approvals; |
| ∎ | product seizure or detention, or refusal to permit the import or export of products; |
| ∎ | consent decrees, corporate integrity agreements, debarment or exclusion from federal healthcare programs; |
| ∎ | mandated modification of promotional materials and labeling and the issuance of corrective information; |
| ∎ | the issuance of safety alerts, Dear Healthcare Provider letters, press releases and other communications containing warnings or other safety information about the product; or |
60
Table of Contents
| ∎ | injunctions or the imposition of fines or civil or criminal penalties. |
Any government investigation of alleged violations of law could require us to expend significant time and resources in response and could generate adverse publicity. Any failure to comply with ongoing regulatory requirements may significantly and adversely affect our ability to commercialize and generate revenue from our products, if approved. If regulatory sanctions are applied or if regulatory approval is withdrawn, the value of our company and our operating results will be adversely affected.
Subsequent discovery of previously unknown problems with a product, including adverse events of unanticipated severity or frequency, or with our CMOs or manufacturing processes, or failure to comply with regulatory requirements, may result in, these same consequences.
Our programs for which we intend to seek approval as biologics may face competition sooner than anticipated.
The Patient Protection and Affordable Care Act, as amended by the Health Care and Education Reconciliation Act (collectively, the ACA), includes a subtitle called the Biologics Price Competition and Innovation Act of 2009 (BPCIA), which created an abbreviated approval pathway for biological products that are biosimilar to or interchangeable with an FDA-licensed reference biological product. Biosimilars are biological products licensed under section 351(k) of the Public Health Service Act (PHS Act) relying on the FDA’s findings of safety, purity, and potency for a licensed biologic (Reference Product) submitted pursuant to section 351(a) of the PHS Act. A biosimilar is highly similar to its Reference Product, excluding minor differences in clinically inactive components for which there are no clinically meaningful differences between the proposed biological product and the Reference Product in safety, purity, or potency. Certain biosimilars that have been designated by FDA as interchangeable may be substituted for the Reference Product in accordance with state law. Under the BPCIA, an application for a biosimilar or interchangeable product relying on the Reference Product may not be submitted to the FDA until four years following the date that the Reference Product was first approved by the FDA. In addition, the approval of a biosimilar product relying on the Reference Product may not be made effective by the FDA until 12 years from the date on which the Reference Product was first approved. During this 12-year period of non-patent data exclusivity, another company may still develop and receive approval of a competing version of the Reference Product if the FDA approves a full BLA for the competing product containing the sponsor’s own nonclinical data and data from adequate and well-controlled clinical trials to demonstrate the safety, purity and potency of their product. The law is complex and is still being interpreted and implemented by the FDA. As a result, its ultimate impact, implementation and meaning are subject to uncertainty, and any new policies or processes adopted by the FDA could have a material effect on the future commercial prospects for our biological product candidates.
We believe that any of our current programs we develop that is approved as biologic under a BLA should qualify for the 12-year period of exclusivity. However, there is a risk that this exclusivity could be shortened due to congressional action or otherwise, or that the FDA will not consider our programs to be Reference Products for competing products, potentially creating the opportunity for biosimilar competition sooner than anticipated. Moreover, the extent to which a biosimilar, once approved, will be substituted for any Reference Product in a way that is similar to traditional generic substitution for non-biological products is not yet clear, and will depend on a number of marketplace and regulatory factors that are still developing. The approval of a biosimilar of our current or future product candidates could have a material adverse impact on our business due to increased competition and pricing pressure.
Similar regulatory exclusivity periods outside the United States, including in the EU and the United Kingdom, differ in scope and duration and may be subject to legal or legislative change, which could also permit competition earlier than expected. In the EU, there is a special regime for biosimilars, or biological medicinal products that are similar to a reference medicinal product but that do not meet the definition of a generic medicinal product. For such products, the results of appropriate nonclinical studies or clinical trials must be provided in support of an MAA. Guidelines from the EMA detail the type of quantity of supplementary data to be provided for different types of biological product. In addition, other countries may have different standards in determining similarity to a Reference Product. Any market entry of competing products to our current or future product candidates in these other regions could adversely affect our business in those regions. To the extent that we do not receive any anticipated periods of regulatory exclusivity for our current or future product candidates, it could adversely affect our business, financial condition, results of operations, and prospects.
61
Table of Contents
The policies of the FDA or other regulatory authorities may change, and additional government regulations may be enacted that could prevent, limit or delay regulatory approval of any of ipsoprubart, ELA822 or our future product candidates.
If we are slow or unable to adapt to changes in existing requirements or the adoption of new requirements or policies, or if we are not able to maintain regulatory compliance, we may lose any regulatory approval that we may have obtained and we may not achieve or sustain profitability, which would adversely affect our business. We also cannot predict the likelihood, nature or extent of government regulation that may arise from future legislation or administrative or executive action, either in the United States or abroad.
In addition, three decisions from the U.S. Supreme Court in July 2024 may lead to an increase in litigation against regulatory agencies that could create uncertainty and thus negatively impact our business. The first decision overturned established precedent that required courts to defer to regulatory agencies’ interpretations of ambiguous statutory language. The second decision overturned regulatory agencies’ ability to impose civil penalties in administrative proceedings. The third decision extended the statute of limitations within which entities may challenge agency actions. These cases may result in increased litigation by industry against regulatory agencies and impact how such agencies choose to pursue enforcement and compliance actions. However, the specific, lasting effects of these decisions, which may vary within different judicial districts and circuits, is unknown. We also cannot predict the extent to which regulations, policies, and decisions of the FDA or other regulatory authorities, such as the SEC, may become subject to increasing legal challenges, delays, and changes.
Moreover, the UK government has enacted new legislation to overhaul the clinical trials regulatory framework. In April 2025, the UK adopted the Medicines for Human Use (Clinical Trials) (Amendment) Regulations 2025, intended to support a more streamlined and flexible regulation of clinical trials, remove unnecessary administrative burdens on trial sponsors, and protect the interests of trial participants. The amendment became applicable on April 28, 2026 following a one-year transition period. While these changes introduce efficiencies, the new requirements may impose additional compliance burdens on us and our CROs conducting trials in the UK and could result in increased costs or delays in trial initiation or conduct.
Furthermore, on December 11, 2025, the European Commission, the European Parliament and the European Council reached a political agreement on a comprehensive overhaul of EU pharmaceutical legislation (the Pharma Package). If formally adopted in its proposed form, the Pharma Package could, among other changes, reduce the baseline market protection period by one year, reshape the incentives regime for orphan medicinal products (including by reducing the baseline orphan market exclusivity from ten to nine years and replacing the current indication-based system with a product-based approach under which subsequent orphan indications for the same active substance will no longer benefit from separate periods of market exclusivity), and expand the Bolar exemption. Under the agreed transitional provisions, marketing authorizations already granted, and MAA submitted before the new regulation becomes applicable, will generally continue to be governed by current protection rules; however, any subsequent application for an additional orphan indication we submit after the entry into force but before the application date, any MAA we submit after the application date, and the expanded Bolar exemption (for which no transitional protection applies) could reduce the period of market exclusivity available to our product candidates in the EU and accelerate exposure to generic or biosimilar competition, potentially materially impacting the commercial prospects of our product candidates. The new framework is expected to enter into force at the end of 2026 to early 2027 and to be subject to transitional arrangements, with full application not anticipated before the end of 2028 to early 2029.
We ultimately cannot predict the likelihood, nature or extent of government regulation that may arise from future legislation or administrative action, either in the United States or abroad. If we are slow or unable to adapt to changes in existing requirements or the adoption of new requirements or policies, or if we are not able to maintain regulatory compliance, we may be subject to enforcement action and our business, results of operations, and financial condition could be adversely affected.
Disruptions at the FDA and other government agencies caused by, among other factors, funding shortages or global health concerns could hinder their ability to hire and retain key leadership and other personnel, slow the time necessary for new products to be reviewed and/or approved, or otherwise prevent those agencies from performing
62
Table of Contents
normal business functions on which the operation of our business may rely, which would adversely affect our business.
The ability of the FDA and foreign regulatory authorities to review and approve new products can be affected by a variety of factors, including government budget and funding levels, government shutdowns, statutory, regulatory, and policy changes, the FDA’s and foreign regulatory authorities’ ability to hire and retain key personnel and accept the payment of user fees, and other events that may otherwise affect the FDA’s and foreign regulatory authorities’ ability to perform routine functions. In addition, government funding of other government agencies on which our operations may rely, including those that fund research and development activities, is subject to the political process, which is inherently fluid and unpredictable.
Disruptions at the FDA and foreign regulatory authorities may slow the time necessary for new products to be reviewed and/or approved, which would adversely affect our business. For example, starting in January 2025, the U.S. government has reduced the number of federal employees, including at FDA, by establishing voluntary termination programs, by position eliminations or by involuntary terminations. Changes in FDA staffing could result in delays in the FDA’s responsiveness or in its ability to review submissions or applications, issue regulations or guidance, or implement or enforce regulatory requirements in a timely fashion or at all.
Similar consequences would also result in the event of a significant shutdown of the federal government. For example, over the last several years, and most recently in late 2025, the U.S. government has shut down several times, and certain regulatory agencies, such as the FDA, had to furlough critical employees and stop critical activities. In addition, the current U.S. Presidential administration has issued certain policies and Executive Orders directed towards reducing the employee headcount and costs associated with U.S. administrative agencies, including the FDA, which have led to substantial personnel changes, and it remains unclear the degree to which these efforts or the resulting changes in FDA personnel may limit or otherwise adversely affect the FDA’s ability to conduct routine activities. If a prolonged government shutdown occurs, or if funding shortages, staffing limitations, or similar factors prevent or hinder the FDA from conducting their regular inspections, reviews, or other regulatory activities, such events could significantly impact the ability of the FDA to timely review and process our regulatory submissions or otherwise provide us with advice or guidance in a timely manner, which could have a material adverse effect on our business. Further, future government shutdowns or delays could impact our ability to access the public markets and obtain necessary capital in order to properly capitalize and continue our operations. If any legislation, executive orders, or lapses in agency funding impose constraints on the FDA’s ability to engage in oversight and implementation activities in the normal course, our business may be negatively impacted.
If we are found to have improperly promoted off-label use of our products, we may become subject to significant liability.
The FDA, comparable foreign regulatory authorities in other jurisdictions strictly regulate the promotional claims that may be made about prescription drug products, such as our products. While physicians, in the practice of medicine, may prescribe approved drugs for unapproved indications, a product may not be promoted for uses that are not approved by the applicable regulatory authority as reflected in the product’s approved labeling or for uses inconsistent with the product’s approved labeling. If our promotional materials and related activities are not consistent with the approved labeling or if physicians, in their professional medical judgment, nevertheless prescribe the drug product to their patients in a manner that is inconsistent with the approved labeling, we may be subject to claims that we promoted off-label use or otherwise violated applicable regulations. In addition, although we may believe ipsoprubart, ELA822 or our future product candidates may provide for superior efficacy as compared to marketed products, without head-to-head data, we will be unable to make comparative claims for our products. If we are found to have promoted such off-label use or made such unsubstantiated comparative claims, we may become subject to significant liability under the Federal Food, Drug, and Cosmetic Act and other statutory authorities, such as laws prohibiting false claims for reimbursement as well as sanctions, fines or other enforcement action under applicable foreign promotional rules and regulations.
Our operations and relationships with healthcare providers, healthcare organizations, customers and third-party payors will be subject to applicable anti-bribery, anti-kickback, fraud and abuse, transparency and other healthcare laws and regulations, which could expose us to, among other things, enforcement actions, criminal sanctions, civil
63
Table of Contents
penalties, contractual damages, reputational harm, administrative burdens and diminished profits and future earnings.
Our current and future arrangements with healthcare providers, healthcare organizations, third- party payors and customers expose us to broadly applicable anti-bribery, fraud and abuse and other healthcare laws and regulations that may constrain the business or financial arrangements and relationships through which we research, market, sell and distribute any of our product candidates, if approved. Restrictions under applicable federal and state anti-bribery and healthcare laws and regulations, include the following:
| ∎ | the federal Anti-Kickback Statute, which prohibits, among other things, individuals and entities from knowingly and willfully soliciting, receiving, offering, or providing remuneration, directly or indirectly, in cash or in kind, to induce or reward, or in return for, either the referral of an individual, or the purchase, order or recommendation of, any good or service for which payment may be made under a federal and state healthcare program such as Medicare and Medicaid. The term remuneration has been broadly interpreted to include anything of value. A person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation; |
| ∎ | the federal criminal and civil false claims and civil monetary penalties laws, including the federal False Claims Act, which can be enforced through civil whistleblower or qui tam actions against individuals or entities, and the Federal Civil Monetary Penalties Law, which prohibit, among other things, knowingly presenting, or causing to be presented, to the federal government, claims for payment that are false or fraudulent, knowingly making, using or causing to be made or used, a false record or statement material to a false or fraudulent claim, or from knowingly making a false statement to avoid, decrease or conceal an obligation to pay money to the federal government. In addition, certain marketing practices, including off-label promotion, may also violate false claims laws. Moreover, the government may assert that a claim including items and services resulting from a violation of the federal Anti-Kickback Statute constitutes a false or fraudulent claim for purposes of the federal False Claims Act; |
| ∎ | HIPAA and its implementing regulations, which imposes criminal and civil liability, prohibits, among other things, knowingly and willfully executing, or attempting to execute a scheme to defraud any healthcare benefit program, or knowingly and willfully falsifying, concealing or covering up a material fact or making any materially false statement in connection with the delivery of or payment for healthcare benefits, items or services; similar to the federal Anti- Kickback Statute, a person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation; |
| ∎ | HIPAA, as amended by HITECH, and their respective implementing regulations, which impose obligations on certain healthcare providers, health plans, and healthcare clearinghouses, known as covered entities, as well as their business associates and covered subcontractors that perform certain services involving the storage, use or disclosure of individually identifiable health information for or on behalf of a covered entity and their business associates, including mandatory contractual terms, with respect to safeguarding the privacy, security, and transmission of individually identifiable health information, and require notification to affected individuals and regulatory authorities of certain breaches of security of individually identifiable health information; |
| ∎ | the federal Physician Payments Sunshine Act, which requires certain manufacturers of covered drugs, devices, biologics and medical supplies that are reimbursable under Medicare, Medicaid, or the Children’s Health Insurance Program, with certain exceptions, to report annually to the Centers for Medicare & Medicaid Services (CMS) information related to certain payments and other transfers of value to physicians (defined to include doctors, dentists, optometrists, podiatrists and chiropractors), certain other health care professionals (such as physician assistants and certain advance practices nurses), and teaching hospitals, as well as ownership and investment interests held by the physicians described above and their immediate family members, with the information made publicly available on a searchable website; |
| ∎ | the Foreign Corrupt Practices Act (FCPA) which prohibits U.S. businesses and their representatives from directly or indirectly offering to pay, paying, promising to pay or authorizing the payment of money or anything of value to a foreign official in order to influence any act or decision of the foreign official in his or her official capacity or to secure any other improper advantage in order to obtain or retain business; |
64
Table of Contents
| ∎ | analogous state and foreign laws and regulations, such as state anti-kickback and false claims laws, that may apply to sales or marketing arrangements and claims involving healthcare items or services reimbursed by non-governmental third-party payors, including private insurers; |
| ∎ | state laws that require the registration of manufacturers and wholesale distributors of drug and biological products who ship into a state, including in certain states that require registration even if such manufacturers or distributors have no place of business within the state. Some states also impose requirements on manufacturers and distributors to establish the pedigree of product in the chain of distribution, including some states that require manufacturers and others to adopt new technology capable of tracking and tracing product as it moves through the distribution chain; |
| ∎ | certain state laws that require biopharmaceutical companies to comply with the biopharmaceutical industry’s voluntary compliance guidelines and the relevant compliance guidance promulgated by the federal government in addition to requiring drug manufacturers to report information related to payments to physicians and other healthcare providers or marketing expenditures and drug pricing information, and state and local laws that require the registration of biopharmaceutical sales representatives; |
| ∎ | analogous foreign laws to the rules described above; and |
| ∎ | foreign as well as U.S. federal and state laws and regulations governing the privacy and security of health information in some circumstances, many of which differ from each other in significant ways and often are not preempted by HIPAA, thus complicating compliance efforts. |
Efforts to ensure that our current and future business arrangements with third parties comply with applicable healthcare laws and regulations could involve substantial costs. It is possible that governmental authorities will conclude that our business practices do not comply with current or future statutes, regulations, agency guidance or case law involving applicable fraud and abuse or other healthcare laws and regulations. If our operations are found to be in violation of any such requirements, we may be subject to significant penalties, including civil, criminal and administrative penalties, damages, fines, disgorgement, imprisonment, the curtailment or restructuring of our operations, loss of eligibility to obtain approvals from the FDA, exclusion from participation in government contracting, healthcare reimbursement or other government programs, including Medicare and Medicaid, integrity oversight and reporting obligations, or reputational harm, any of which could adversely affect our financial results. These risks cannot be entirely eliminated. Any action against us for an alleged or suspected violation could cause us to incur significant legal expenses and could divert our management’s attention from the operation of our business, even if our defense is successful. In addition, achieving and sustaining compliance with applicable laws and regulations may be costly to us in terms of money, time and resources.
We may face difficulties from healthcare legislative and regulatory reform measures.
Existing laws and regulatory policies may change and additional government regulations may be enacted that could prevent, limit or delay regulatory approval of ipsoprubart, ELA822 or any of our future product candidates. We cannot predict the likelihood, nature or extent of government regulation that may arise from future legislation or administrative action, either in the United States or abroad. If we are slow or unable to adapt to changes in existing requirements or the adoption of new requirements or policies, or if we are not able to maintain regulatory compliance, we may lose any regulatory approval that we may have obtained, or may face penalties for any approved products, and we may not achieve or sustain profitability.
In the United States and some foreign jurisdictions, there have been and we expect there will continue to be, a number of legislative and regulatory changes to the healthcare system, including cost containment measures that may reduce or limit coverage and reimbursement for newly approved drugs and affect our ability to profitably sell any product candidates for which we obtain regulatory approval. In particular, there have been and continue to be a number of initiatives at the U.S. federal and state levels that seek to reduce healthcare costs and improve the quality of care.
For example, the ACA made significant changes to the healthcare system in the United States. Since its enactment, there have been amendments and judicial, Congressional and executive branch challenges to certain aspects of the ACA. For example, on July 4, 2025, the OBBBA was signed into law, which narrowed access to ACA marketplace exchange enrollment and declined to extend the ACA enhanced advanced premium tax credits that expired at the
65
Table of Contents
end of 2025, which, among other provisions in the law, are anticipated to reduce the number of Americans with health insurance. The OBBBA also is expected to reduce Medicaid spending and enrollment by implementing work requirements for some beneficiaries, capping state-directed payments, reducing federal funding, and limiting provider taxes used to fund the program. Congress is considering proposed legislation intended to further reduce healthcare costs with alternatives to replace the expired ACA subsidies.
Other legislative changes have been proposed and adopted in the United States since the ACA was enacted. These changes include aggregate reductions to Medicare payments to providers, which began in 2013 and will remain in effect until 2032 unless additional Congressional action is taken.
The federal administration is pursuing policies to reduce regulations and expenditures across government, including at the U.S. Department of Health and Human Services (HHS), which include the FDA and CMS, and related agencies. These actions, presently directed by executive orders or memoranda from the Office of Management and Budget, may propose policy changes that create additional uncertainty for our business. For example, the current administration has announced agreements with several pharmaceutical companies that require the drug manufacturers to offer, through a direct to consumer platform (TrumpRx) U.S. patients and Medicaid programs prescription drug Most-Favored Nation pricing equal to or lower than those paid in other developed nations, with additional mandates for direct-to-patient discounts and repatriation of foreign revenues. Other recent actions, for example, include (1) directing agencies to reduce agency workforce and cut programs; (2) directing HHS and other agencies to lower prescription drug costs through a variety of initiatives; (3) imposing tariffs on certain imported pharmaceutical products; and (4) as part of the Make America Healthy Again Commission’s Strategy Report released in September 2025, working across government agencies to increase enforcement on direct-to-consumer pharmaceutical advertising. Additionally, the current administration recently called on Congress to enact “The Great Healthcare Plan,” to codify and expand Most-Favored Nation pricing, lower government subsidies to private insurance companies, increase healthcare price transparency, expand pharmaceutical drugs available for over-the-counter purchase, and enact restrictions on pharmacy benefit manager payment methodologies, among other things.
Further, in its June 2024 decision in Loper Bright Enterprises v. Raimondo, the U.S. Supreme Court overturned the longstanding Chevron doctrine, under which courts were required to give deference to regulatory agencies’ reasonable interpretations of ambiguous federal statutes. The Loper Bright decision could result in additional legal challenges to current regulations and guidance issued by federal agencies applicable to our operations, including those issued by the FDA.
At the state level, legislatures have increasingly passed legislation and implemented regulations designed to control pharmaceutical and biological product pricing, including price or patient reimbursement constraints, discounts, restrictions on certain product access, marketing cost disclosure, drug price reporting and other transparency measures. Some states have enacted legislation creating so-called prescription drug affordability boards, which ultimately may attempt to impose price limits on certain drugs in these states, while some states are also seeking to implement general, across the board price caps for pharmaceuticals, or are seeking to regulate drug distribution. Some measures are designed to encourage importation from other countries and bulk purchasing. Legally mandated price controls on payment amounts by third-party payors or other restrictions could seriously harm our business. In addition, regional healthcare authorities and individual hospitals are increasingly using bidding procedures to determine what pharmaceutical products and which suppliers will be included in their prescription drug and other healthcare programs. This could reduce the ultimate demand for our product candidates or put pressure on our product pricing. Furthermore, there has been increased interest by third-party payors and governmental authorities in reference pricing systems and publication of discounts and list prices. Prescription drugs and biological products that are in violation of these requirements will be included on a public list. These reforms could reduce the ultimate demand for our product candidates or put pressure on our product pricing and could seriously harm our business.
We expect that additional state, federal and foreign healthcare reform measures will be adopted in the future, any of which could limit the amounts that federal and state governments will pay for pharmaceuticals and other healthcare products and services, which could result in reduced demand for ipsoprubart, ELA822 or any of our future product candidates or additional pricing pressures or otherwise impact our operations.
66
Table of Contents
Even if we are able to commercialize ipsoprubart, ELA822 and any future product candidates, if approved, such product candidate may become subject to unfavorable pricing regulations or third-party coverage and reimbursement policies, which would harm our business.
The availability of coverage and the adequacy of reimbursement by governmental healthcare programs, such as Medicare and Medicaid, private health insurers and other third-party payors are essential for most patients to be able to afford prescription medications such as our product candidates, if approved. Sales of any of our product candidates that receive regulatory approval will be dependent substantially, both in the United States and internationally, on the extent to which the costs of our product candidates will be paid by health maintenance, managed care, pharmacy benefit, and similar healthcare management organizations or reimbursed by government health administration authorities, private health coverage insurers and other third-party payors. Even if coverage is provided, the approved reimbursement amount may not be high enough to allow us to establish or maintain pricing sufficient to realize an adequate return on our investment. If reimbursement is not available, or is available only to limited levels, we may not be able to successfully commercialize our product candidates. Coverage and reimbursement may impact the demand for, or the price of, any product candidate for which we obtain regulatory approval. If coverage and reimbursement are not available or reimbursement is available only to limited levels, we may not successfully commercialize any product candidate for which we obtain regulatory approval.
There is significant uncertainty related to insurance coverage and reimbursement of newly approved products. In the United States, private payors often follow the coverage and reimbursement policies established by CMS, the federal agency responsible for administering the Medicare program. One payor’s determination to provide coverage for a product does not assure that other payors will also provide coverage for the product. As a result, the coverage determination process is often time-consuming and costly. This process will require us to provide scientific and clinical support for the use of our products to each payor separately, with no assurance that coverage and adequate reimbursement will be applied consistently or obtained in the first instance. Increasingly, third-party payors are requiring that drug companies provide them with predetermined discounts from list prices and are challenging the prices charged for medical products. Further, such payors are increasingly challenging the price, examining the medical necessity and reviewing the cost effectiveness of medical products. There may be especially significant delays in obtaining coverage and reimbursement for newly approved products. Third-party payors may limit coverage to specific products on an approved list, known as a formulary, which might not include all FDA-approved products for a particular indication. We may need to conduct expensive pharmaco-economic studies to demonstrate the medical necessity and cost effectiveness of our products. Nonetheless, our product candidates may not be considered medically necessary or cost effective. We cannot be sure that coverage and reimbursement will be available for any product that we commercialize and, if reimbursement is available, what the level of reimbursement will be.
Moreover, increasing efforts by governmental and third-party payors in the United States and abroad to cap or reduce healthcare costs may cause such organizations to limit both coverage and the level of reimbursement for newly approved products and, as a result, they may not cover or provide adequate payment for our product candidates, if approved. For example, HHS imposes rebates on many Medicare Part B and Medicare Part D products to penalize price increases that outpace inflation on an annual basis. HHS has also been empowered to negotiate the price of certain single-source drugs that have been on the market for seven (7) years and single-source biologics that have been on the market for at least eleven (11) years covered under Medicare as part of the Medicare drug price negotiation program. Each year up to twenty (20) products will be selected by HHS for the Medicare drug price negotiation program. Products subject to the Medicare drug price negotiation program are expected to experience a significant reduction in reimbursement from the Medicare program on a per unit basis. We expect to experience pricing pressures in connection with the sale of our product candidates due to the trend toward managed healthcare, the increasing influence of health maintenance organizations and additional legislative changes. The downward pressure on healthcare costs in general, particularly prescription drugs and surgical procedures and other treatments, has become very intense. As a result, increasingly high barriers are being erected to the entry of new products.
We are subject to U.S. and certain foreign export and import controls, sanctions, embargoes, anti-corruption laws and other trade laws and regulations. Compliance with these legal standards could impair our ability to compete in
67
Table of Contents
domestic and international markets. We can face criminal liability and other serious consequences for violations, which can harm our business.
U.S. and foreign anti-corruption laws and regulations prohibit, among other things, companies and their employees, agents, CROs, CMOs, consultants, contractors and other partners and representatives from authorizing, promising, offering, providing, soliciting, or receiving directly or indirectly, corrupt or improper payments or anything else of value to or from recipients in the public or private sector. The FCPA also obligates companies whose securities are listed in the United States to comply with accounting provisions requiring the company to maintain books and records that accurately and fairly reflect all transactions of the corporation, including international subsidiaries, and to devise and maintain an adequate system of internal accounting controls. Violations of these laws can result in substantial criminal fines and civil penalties, imprisonment, the loss of trade privileges, debarment, tax reassessments, breach of contract and fraud litigation, reputational harm and other consequences. We have direct or indirect interactions with officials and employees of government agencies or government-affiliated hospitals, universities and other organizations. We also expect our non-U.S. activities to increase over time. We expect to rely on third parties for research, nonclinical studies and clinical trials and/or to obtain necessary permits, licenses, patent registrations and other regulatory approvals. We can be held liable for the corrupt or other illegal activities of our personnel, agents, or partners, even if we do not explicitly authorize or have prior knowledge of such activities.
We are also subject to export control, import, and sanctions laws and regulations, including the U.S. Export Administration Regulations, U.S. Customs regulations and various economic and trade sanctions regulations administered by the U.S. Treasury Department’s Office of Foreign Assets Control. Compliance with applicable regulatory requirements regarding the export of our products may create delays in the introduction of our products in international markets or, in some cases, prevent the export of our products to some countries altogether. Furthermore, U.S. export control laws and economic sanctions prohibit the provision of certain products and services to countries, governments and persons targeted by U.S. sanctions.
Any violations of the laws and regulations described above may result in substantial civil and criminal fines and penalties, imprisonment, the loss of export or import privileges, debarment, tax reassessments, breach of contract and fraud litigation, reputational harm and other consequences.
Governments outside the United States tend to impose strict price controls, which may adversely affect our revenue, if any.
In some countries, particularly member states of the EU, the pricing of certain therapeutic products is subject to governmental control. In these countries, pricing negotiations with governmental authorities can take considerable time after receipt of regulatory approval for a product. In addition, there can be considerable pressure by governments and other stakeholders on prices and reimbursement levels, including as part of cost containment measures. Political, economic and regulatory developments may further complicate pricing negotiations, and pricing negotiations may continue after reimbursement has been obtained.
In the EU, pricing and reimbursement decisions are made by individual EU Member States and are separate from the marketing authorization process. EU Member States may restrict the range of products for which their national health insurance systems provide reimbursement, control the prices of medicinal products, or impose significant price discounts or rebates. In addition, EU Member States increasingly require health technology assessments (HTAs) that compare the cost-effectiveness of a medicinal product to currently available therapies as a condition of reimbursement. At the EU level, Regulation (EU) 2021/2282 on HTA entered into application on January 12, 2025 through a phased implementation. The HTA Regulation initially applies to new active substances for oncology products and advanced therapy medicinal products, and will expand to orphan medicinal products in January 2028 and to all centrally authorized medicinal products by 2030. As the HTA Regulation applies to orphan medicinal products from 2028, joint clinical assessments conducted at the EU level may result in adverse or delayed outcomes that negatively affect our ability to obtain or maintain favorable reimbursement status across EU Member States. Any failure to obtain adequate coverage and reimbursement in EU Member States for our product candidates could limit our commercial opportunity and materially harm our business, financial condition, and results of operations.
Reference pricing used by various EU member states and parallel distribution, or arbitrage between low-priced and high-priced member states, can further reduce prices. To obtain coverage and reimbursement or pricing approvals in
68
Table of Contents
some countries, we may be required to conduct a clinical trial or other studies that compare the cost-effectiveness of ipsoprubart, ELA822 or any of our future product candidates to other available therapies in order to obtain or maintain reimbursement or pricing approval. Publication of discounts by third-party payors or authorities may lead to further pressure on the prices or reimbursement levels within the country of publication and other countries. If reimbursement of any product candidate approved for marketing is unavailable or limited in scope or amount, or if pricing is set at unsatisfactory levels, our business, financial condition, results of operations or prospects could be materially and adversely affected.
Risks Related to Our Common Stock and This Offering
The market price of our common stock is likely to be highly volatile, and you could lose all or part of your investment.
The trading price of our common stock following this offering is likely to be highly volatile and subject to wide fluctuations in response to various factors, some of which we cannot control. As a result of this volatility, investors may not be able to sell their common stock at or above the initial public offering price. The market price for our common stock may be influenced by many factors, including the other risks described in this “Risk Factors” section and the following:
| ∎ | results of nonclinical studies and clinical trials of any product candidates, or those of our competitors or any potential future collaborators or licensing partners; |
| ∎ | the timing and enrollment status of our clinical trials; |
| ∎ | regulatory or legal developments in the United States or other countries, especially changes in federal or global health policies, laws or regulations applicable to any product candidates, including the review and oversight functions of federal health regulatory bodies; |
| ∎ | the success or failure of competitive products or technologies; |
| ∎ | introductions and announcements of new product candidates by us, any future commercialization partners, or our competitors, and the timing of these introductions or announcements; |
| ∎ | actions taken by regulatory agencies with respect to any product candidates, clinical trials, and, if approved, manufacturing process or sales and marketing terms; |
| ∎ | actual or anticipated variations in our financial results or those of companies that are perceived to be similar to us; |
| ∎ | the success of our efforts to identify, acquire or in-license new technologies or product candidates; |
| ∎ | developments concerning any future collaborations, including but not limited to those with development and commercialization partners if any product candidates are approved; |
| ∎ | market conditions in the pharmaceutical and biotechnology sectors; |
| ∎ | announcements by us or our competitors of significant acquisitions, strategic collaborations, joint ventures or capital commitments; |
| ∎ | developments or disputes concerning patents or other proprietary rights, including patents, litigation matters and our ability to obtain patent protection for any product candidates; |
| ∎ | our ability or inability to raise additional capital and the terms on which we are able to raise it, if at all; |
| ∎ | our ability to effectively manage our growth; |
| ∎ | the recruitment or departure of key personnel; |
| ∎ | changes in the structure of healthcare payment systems; |
| ∎ | actual or anticipated changes in earnings estimates, development timelines or changes in stock market analyst recommendations regarding our common stock, other comparable companies or our industry generally; |
| ∎ | our failure or the failure of our competitors to meet analysts’ projections or guidance that we or our competitors may give to the market; |
| ∎ | fluctuations in the valuation of companies perceived by investors to be comparable to us; |
| ∎ | announcement and expectation of additional financing efforts; |
69
Table of Contents
| ∎ | speculation in the press or investment community; |
| ∎ | fluctuations of share price and trading volume of our common stock; |
| ∎ | sales or perceived potential sales of shares of our common stock by us, insiders or our stockholders; |
| ∎ | the concentrated ownership of our common stock; |
| ∎ | expiration of market stand-off or lock-up agreements; |
| ∎ | changes in accounting principles; |
| ∎ | actions instituted by activist stockholders or others; |
| ∎ | terrorist acts, acts of war or periods of widespread civil unrest; |
| ∎ | natural disasters and other calamities, including global epidemics and pandemics such as the COVID-19 pandemic; |
| ∎ | general economic, industry and market conditions, including changes in tariffs and trade restrictions, fluctuating interest rates and inflation; and |
| ∎ | other events or factors, many of which are beyond our control. |
In addition, the stock market in general, and the markets for pharmaceutical, biopharmaceutical and biotechnology stocks in particular, have experienced extreme price and volume fluctuations that have been often unrelated or disproportionate to the operating performance of the issuer. Furthermore, the trading price of our common stock may be adversely affected by third parties trying to drive down the market price. Short sellers and others, some of whom post anonymously on social media, may be positioned to profit if our stock declines and their activities can negatively affect our stock price. These broad market and industry factors may seriously harm the market price of our common stock, regardless of our actual operating performance. The realization of any of the above risks or any of a broad range of other risks, including those described in this “Risk Factors” section, could have a dramatic and adverse impact on the market price of our common stock.
Sales of substantial amounts of shares of our common stock may cause the price of our common stock to decline.
Based on 39,433,621 shares of our capital stock outstanding as of June 30, 2026, assuming the automatic conversion of our outstanding redeemable convertible preferred stock as of June 30, 2026, upon completion of this offering, we will have a total of 61,100,288 shares of common stock outstanding. Of these shares, only the 21,666,667 shares of common stock sold in this offering, or 24,916,667 shares if the underwriters exercise their option to purchase additional shares in full, will be freely tradable, without restriction, in the public market immediately after this offering. Each of our officers, directors and substantially all of our stockholders have entered into lock-up agreements with the underwriters that, among other things and subject to certain exceptions, restrict their ability to sell or transfer their shares. The lock-up agreements pertaining to this offering will expire 180 days from the date of this prospectus. However, Jefferies LLC and TD Securities (USA) LLC may, in their sole discretion, permit our officers, directors and other stockholders who are subject to the lock-up agreements to sell shares prior to the expiration of the lock-up agreements. After the lock-up agreements expire, based on 39,433,621 shares outstanding as of June 30, 2026, assuming the automatic conversion of our outstanding redeemable convertible preferred stock as of June 30, 2026, approximately all 39,433,621 shares of common stock will be eligible for sale in the public market, of which approximately 14,718,264 shares are held by our officers, directors and their affiliated entities, which assumes that Electra Therapeutics LLC (Electra LLC) uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock and will be subject to volume limitations under Rule 144 under the Securities Act of 1933, as amended (Securities Act). See the section titled “Executive and Director Compensation—Narrative to Summary Compensation Table—Equity-based Incentive Awards” for additional information about Electra LLC. Prior to the closing of this offering, Electra LLC expects to distribute shares of our Series A redeemable convertible preferred stock to the unit holders of Electra LLC, which includes certain of our officers, directors and their affiliated entities.
After this offering, the holders of an aggregate of 39,404,292 shares of our outstanding common stock as of June 30, 2026 will have rights, subject to some conditions, to require us to file registration statements covering their shares or to include their shares in registration statements that we may file for ourselves or our stockholders. We also intend to register shares of common stock that we may issue under our equity incentive plans. Once we register these shares, they will be able to be sold freely in the public market upon issuance, subject to the 180-day lock-up period under
70
Table of Contents
the lock-up agreements described above and in the sections titled “Shares Eligible for Future Sale” and “Underwriting.” See the section titled “Description of Capital Stock—Registration Rights” for additional information.
We cannot predict what effect, if any, sales of our shares in the public market or the availability of shares for sale will have on the market price of our common stock. However, future sales of substantial amounts of our common stock in the public market, including shares issued upon exercise of our outstanding options, or the perception that such sales may occur, could adversely affect the market price of our common stock.
We also expect that significant additional capital may be needed in the future to continue our planned operations. To raise capital, we may sell common stock, convertible securities or other equity securities in one or more transactions at prices and in a manner we determine from time to time. To the extent that additional capital is raised through the sale and issuance of shares of our common stock or other securities convertible into shares of our common stock, our stockholders will be diluted. These sales, or the perception in the market that the holders of a large number of shares intend to sell shares of our common stock, could reduce the market price of our common stock.
We will have broad discretion in the use of the net proceeds from this offering and may not use them effectively.
Our management will have broad discretion in the application of the net proceeds from this offering, including for any of the purposes described in the section titled “Use of Proceeds,” and you will be relying on the judgment of our management regarding the application of these proceeds. You will not have the opportunity, as part of your investment decision, to assess whether we are using the proceeds appropriately. Our management might not apply our net proceeds in ways that ultimately increase the value of your investment. If we do not invest or apply the net proceeds from this offering in ways that enhance stockholder value, we may fail to achieve expected financial results, which could cause our stock price to decline. Pending their use, we may invest the net proceeds from this offering in a manner that does not produce income or that loses value.
No public market for our common stock currently exists, and an active and liquid trading market for our common stock may never develop. As a result, you may not be able to resell your shares of common stock at or above the initial public offering price.
Prior to this offering, no market for our common stock existed and an active trading market for our common stock may never develop or be sustained following this offering. The initial public offering price for our common stock was determined through negotiations with the underwriters and the negotiated price may not be indicative of the market price of our common stock after this offering. This price will not necessarily reflect the price at which investors in the market will be willing to buy and sell our shares following this offering, and the market value of our common stock may decrease from the initial public offering price. As a result of these and other factors, you may be unable to resell your shares of our common stock at or above the initial public offering price. The lack of an active market may impair your ability to sell your shares of common stock at the time you wish to sell them or at a price that you consider reasonable. The lack of an active market may also reduce the fair market value of your shares of common stock. To the extent certain of our existing stockholders and their affiliated entities participate in this offering, such purchases would reduce the nonaffiliated public float of our shares, meaning the number of shares of our common stock that are not held by officers, directors and controlling stockholders. As a result, the number of freely tradeable shares of our common stock following this offering will be reduced to what it would have been had these shares been sold to investors that were not existing stockholders, affiliates or purchasers. A reduction in the public float could reduce the number of shares that are available to be traded at any given time, thereby adversely impacting the liquidity of our common stock and depressing the price at which you may be able to sell shares of common stock purchased in this offering. Furthermore, an inactive market may also impair our ability to raise capital by selling shares of our common stock and may impair our ability to enter into strategic collaborations or acquire companies or products by using our shares of common stock as consideration.
You will experience immediate and substantial dilution as a result of this offering and raising additional capital in the future may cause dilution to our stockholders, including purchasers of common stock in this offering, restrict our operations or require us to relinquish rights to our technologies or product candidates.
The assumed initial public offering price of $15.00 per share, which is the midpoint of the range set forth on the cover page of this prospectus, is substantially higher than the pro forma as adjusted net tangible book value per share of our outstanding common stock immediately following the completion of this offering. If you purchase common stock in this offering at the assumed initial public offering price of $15.00 per share, and assuming that
71
Table of Contents
the underwriters do not exercise their option to purchase additional common stock in this offering, you will incur immediate and substantial dilution of $8.68 per share, representing the difference between the assumed initial public offering price of $15.00 per share and our pro forma as adjusted net tangible book value per share as of June 30, 2026 after giving effect to this offering and the conversion of all outstanding redeemable convertible preferred stock immediately prior to the closing of this offering. Following the completion of this offering, investors purchasing common stock in this offering will have contributed 51.9% of the total amount invested by stockholders since inception, but will only own 35.5% of the shares of common stock outstanding. For a further description of the dilution you will experience immediately after this offering, see the section titled “Dilution.”
After this offering, our executive officers, directors and principal stockholders, if they choose to act together, will continue to have the ability to control or significantly influence all matters submitted to stockholders for approval.
Upon the closing of this offering, our executive officers, directors and stockholders who owned more than 5% of our outstanding common stock before this offering and their respective affiliates will, in the aggregate, hold approximately 34.0% of our outstanding common stock (based on the number of shares of common stock outstanding as of June 30, 2026 and after the conversion of all outstanding redeemable convertible preferred stock immediately prior to the closing of this offering, assuming no purchase of shares in this offering by any of this group, and assuming that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock). As a result, if these stockholders choose to act together, they would be able to control or significantly influence all matters submitted to our stockholders for approval, as well as our management and affairs. For example, these persons, if they choose to act together, would control or significantly influence the election of directors, the composition of our management and approval of any merger, consolidation, sale of all or substantially all of our assets or other business combination that other stockholders may desire. The interests of these stockholders may not always coincide with your interests or the interests of other stockholders and they may act in a manner that advances their best interests and not necessarily of those of other stockholders, including seeking a premium value for their common stock. Any of these actions could adversely affect the market price of our common stock.
Anti-takeover provisions in our charter documents and under Delaware law could prevent or delay an acquisition of us, which may be beneficial to our stockholders, and may prevent attempts by our stockholders to replace or remove our current management.
Our amended and restated certificate of incorporation and our amended and restated bylaws that will be in effect upon completion of this offering contain provisions that could delay or prevent a change in control of our company. These provisions could also make it difficult for stockholders to elect directors who are not nominated by current members of our board of directors or take other corporate actions, including effecting changes in our management. These provisions:
| ∎ | establish a classified board of directors so that not all members of our board of directors are elected at one time; |
| ∎ | permit only the board of directors to establish the number of directors and fill vacancies on the board of directors; |
| ∎ | provide that directors may only be removed “for cause” and only with the approval of two-thirds of our stockholders; |
| ∎ | require super-majority voting to amend some provisions in our amended and restated certificate of incorporation and amended and restated bylaws; |
| ∎ | authorize the issuance of “blank check” preferred stock that our board of directors could use to implement a stockholder rights plan; |
| ∎ | eliminate the ability of our stockholders to call special meetings of stockholders; |
| ∎ | prohibit stockholder action by written consent, which requires all stockholder actions to be taken at a meeting of our stockholders; |
| ∎ | prohibit cumulative voting; and |
| ∎ | establish advance notice requirements for nominations for election to our board of directors or for proposing matters that can be acted upon by stockholders at annual stockholder meetings. |
In addition, Section 203 of the General Corporation Law of the State of Delaware (DGCL), may discourage, delay or prevent a change in control of our company. Section 203 imposes certain restrictions on mergers, business combinations and other transactions between us and holders of 15% or more of our common stock.
72
Table of Contents
The exclusive forum provisions in our organizational documents may limit a stockholder’s ability to bring a claim in a judicial forum that it finds favorable for disputes with us or any of our directors, officers, or employees, or the underwriters of any offering giving rise to such claim, which may discourage lawsuits with respect to such claims.
Our amended and restated bylaws that will be in effect upon completion of this offering, to the fullest extent permitted by law, will provide that the Court of Chancery of the State of Delaware is the exclusive forum for: any derivative action or proceeding brought on our behalf; any action asserting a breach of fiduciary duty; any action asserting a claim against us arising pursuant to the DGCL, our amended and restated certificate of incorporation, or our amended and restated bylaws; or any action asserting a claim that is governed by the internal affairs doctrine. This exclusive forum provision does not apply to suits brought to enforce a duty or liability created by the Exchange Act. It could apply, however, to a suit that falls within one or more of the categories enumerated in the exclusive forum provision. This choice of forum provision may limit a stockholder’s ability to bring a claim in a judicial forum that it finds favorable for disputes with us or any of our directors, officers, or other employees, or the underwriters of any offering giving rise to such claims, which may discourage lawsuits with respect to such claims. Alternatively, if a court were to find the choice of forum provisions contained in our amended and restated bylaws to be inapplicable or unenforceable in an action, we may incur additional costs associated with resolving such action in other jurisdictions, which could harm our business, financial condition, results of operations, and prospects.
Section 22 of the Securities Act creates concurrent jurisdiction for federal and state courts over all claims brought to enforce any duty or liability created by the Securities Act or the rules and regulations thereunder. Our amended and restated bylaws will provide that the federal district courts of the United States will, to the fullest extent permitted by law, be the exclusive forum for resolving any complaint asserting a cause of action arising under the Securities Act (Federal Forum Provision). Our decision to adopt a Federal Forum Provision followed a decision by the Supreme Court of the State of Delaware holding that such provisions are facially valid under Delaware law. While federal or other state courts may not follow the holding of the Delaware Supreme Court or may determine that the Federal Forum Provision should be enforced in a particular case, application of the Federal Forum Provision means that suits brought by our stockholders to enforce any duty or liability created by the Securities Act must be brought in federal court and cannot be brought in state court, and our stockholders cannot waive compliance with the federal securities laws and the rules and regulations thereunder. Section 27 of the Exchange Act creates exclusive federal jurisdiction over all claims brought to enforce any duty or liability created by the Exchange Act or the rules and regulations thereunder. In addition, neither the exclusive forum provision nor the Federal Forum Provision applies to suits brought to enforce any duty or liability created by the Exchange Act. Accordingly, actions by our stockholders to enforce any duty or liability created by the Exchange Act or the rules and regulations thereunder must be brought in federal court, and our stockholders cannot waive compliance with the federal securities laws and the rules and regulations thereunder.
Any person or entity purchasing or otherwise acquiring or holding any interest in any of our securities shall be deemed to have notice of and consented to our exclusive forum provisions, including the Federal Forum Provision. These provisions may limit a stockholders’ ability to bring a claim, and may result in increased costs for a stockholder to bring such a claim, in a judicial forum of their choosing for disputes with us or our directors, officers, other employees or agents, which may discourage lawsuits against us and our directors, officers, other employees or agents.
We do not currently intend to pay dividends on our common stock and, consequently, our stockholders’ ability to achieve a return on their investment will be dependent on appreciation of the value of our common stock.
We have never declared or paid any cash dividends on our common stock. We currently intend to retain all available funds and any future earnings to support operations and to finance the growth and development of our business. As a result, any investment return on our common stock will be dependent on increases in the value for our common stock, which is not certain. There is no guarantee that shares of our common stock will appreciate in value or even maintain the price at which stockholders have purchased their shares.
If securities or industry analysts do not publish research or reports about our business, or if they issue an adverse or misleading opinion regarding our stock, our stock price and trading volume could decline.
The trading market for our common stock will be influenced by the research and reports that industry or securities analysts publish about us or our business. We do not have any control over the industry or securities analysts, or the
73
Table of Contents
content and opinions included in their reports. We do not currently have and may never obtain research coverage by securities and industry analysts. If no or few securities or industry analysts commence coverage of us, the trading price for our common stock could be impacted negatively. In the event we obtain securities or industry analyst coverage, if any of the analysts who cover us issue an adverse or misleading opinion regarding us, our business model, our intellectual property or our stock performance, or if our nonclinical studies and clinical trials and operating results fail to meet the expectations of analysts, our stock price would likely decline. If one or more of such analysts cease coverage of us or fail to publish reports on us regularly, we could lose visibility in the financial markets, which in turn could cause a decline in our stock price or trading volume.
General Risk Factors
Our current in-person operations are located in California, and we or the third parties on whom we depend may be adversely affected by natural disasters, terrorist activity, pandemics, geo-political actions in the United States and in foreign countries, and other events beyond our control, and our business continuity and disaster recovery plans may not adequately protect us from a serious disaster. Geo-political actions could increase the uncertainties and costs surrounding the prosecution or maintenance of our patent applications or those of any current or future licensors and the maintenance, enforcement or defense of our issued patents or those of any current or future licensors.
Our in-person operations are located in our corporate headquarters and research and development facility in South San Francisco, California. Any unplanned event, such as flood, fire, explosion, earthquake, extreme weather condition, pandemic, medical epidemic, power shortage, telecommunication failure or other natural or manmade accidents or incidents that result in us being unable to fully utilize our facilities, or the manufacturing facilities of our CMOs may have a material and adverse effect on our ability to operate our business and have significant negative consequences on our financial and operating conditions. If our facilities, or the manufacturing facilities of our CMOs, are unable to operate because of an accident or incident or for any other reason, including an inability to use all or a significant portion of our headquarters, damages to critical infrastructure, such as our research facilities or the manufacturing facilities of our CMOs, or other disruptions to operations, even for a short period of time, any or all of our research and development programs may be harmed. Any business interruption could have a material and adverse effect on our business, financial condition, results of operations, and prospects.
Our employees often conduct business outside of any facilities leased by us. These locations may be subject to additional security and other risk factors due to the limited control of our employees. The disaster recovery and business continuity plans we have in place may prove inadequate in the event of a serious disaster or similar event. We may incur substantial expenses as a result of the limited nature of our disaster recovery and business continuity plans, which could have a material adverse effect on our business. As part of our risk management policy, we maintain insurance coverage at levels that we believe are appropriate for our business. However, in the event of an accident or incident at these facilities, we cannot assure you that the amounts of insurance will be sufficient to satisfy any damages and losses.
Unstable market and economic conditions and adverse developments affecting the financial services industry, such as actual events or concerns involving inflation, liquidity, defaults or nonperformance by financial institutions or transactional counterparties, could adversely affect our current and projected business operations, and our financial condition and results of operations.
From time to time, the global credit and financial markets have experienced extreme volatility and disruptions, including severely diminished liquidity and credit availability, declines in consumer confidence, declines in economic growth, increases in unemployment rates and uncertainty about economic stability. The financial markets and the global economy may also be adversely affected by the current or anticipated impact of military conflict, terrorism or other geopolitical events. Sanctions imposed by the United States and other countries in response to such conflicts, including the ongoing conflict in Ukraine and ongoing conflicts in the Middle East, which has recently included actions by Iran, Israel and the United States, may also adversely impact the financial markets and the global economy, and any economic countermeasures by the affected countries or others could exacerbate market and economic instability. Russia’s ongoing incursion of Ukraine has created extreme volatility in the global capital markets and is expected to have further global economic consequences, including disruptions of the global supply chain and energy markets; it is possible that ongoing conflicts in the Middle East may have similar effects. In
74
Table of Contents
addition, adverse developments that affect financial institutions, such as events involving liquidity that are rumored or actual, have in the past and may in the future lead to market-wide liquidity problems. For example, in March 2023, Silicon Valley Bank (SVB), one of our banking partners, was closed by the California Department of Financial Protection and Innovation, which appointed the Federal Deposit Insurance Corporation (FDIC) as receiver. The closure of any additional national or regional commercial banks could lead to further economic instability. Although the Department of the Treasury, the Federal Reserve and the FDIC have taken steps to mitigate these risks, widespread demands for customer withdrawals or other liquidity needs of financial institutions for immediate liquidity may still occur in the future. We may maintain cash balances at third-party financial institutions in excess of the FDIC insurance limit and there is no guarantee that the federal government would provide access to uninsured funds in the future in the event of the closure of other banks or financial institutions, or that they would do so in a timely fashion.
Although we have not experienced any adverse impact to our liquidity or to our current and projected business operations, financial condition or results of operations, uncertainty remains over liquidity concerns in the broader financial services industry, and our business, our business partners, or industry as a whole may be adversely impacted in ways that we cannot predict at this time. Inflation and fluctuating interest rates have led to a decline in the trading value of previously issued government securities with interest rates below current market interest rates.
International trade policies, including tariffs, sanctions and trade barriers may adversely affect our business, financial condition, results of operations, and prospects.
We operate in a global economy, which includes utilizing third-party suppliers in several countries outside the United States. There is inherent risk, based on the complex relationships among the U.S. and the countries in which we conduct our business, that political, diplomatic, and national security factors can lead to global trade restrictions and changes in trade policies and export regulations that may adversely affect our business and operations. The current international trade and regulatory environment is subject to significant ongoing uncertainty. The U.S. government has announced substantial new tariffs affecting a wide range of products and jurisdictions and has indicated an intention to continue developing new trade policies, including with respect to the pharmaceutical industry. In response, certain foreign governments have announced or implemented retaliatory tariffs and other protectionist measures. These developments have created a dynamic and unpredictable trade landscape, which may adversely impact our business, results of operations, financial condition and prospects. The Bureau of Industry and Security, U.S. Department of Commerce, has initiated an investigation to determine whether pharmaceutical ingredients, including finished drug product, manufactured outside the United States pose a national security risk and should be subject to additional tariffs. Following that investigation, the President announced a proclamation which will impose 100% tariffs on certain patented pharmaceutical products and associated pharmaceutical ingredients. We are assessing the potential impact of the proclamation on our business.
We do not own or operate, and currently have no plans to establish, any manufacturing facilities. We currently rely, and expect to continue to rely, on third parties for the manufacture of our product candidates for clinical testing, as well as for manufacture of any products that we may commercialize, if approved. Currently, several of our suppliers are located outside of the United States, and our principal suppliers of critical raw materials are located in the UK, United States, China, and Ireland. The DS for ipsoprubart is manufactured in the UK and ELA822 DS is manufactured in China. The DP for ipsoprubart is manufactured in the United States and ELA822 DP is manufactured in China. We also rely on specialized laboratory equipment, supplies, materials, and precursor compounds, all or part of which we believe may be ultimately sourced from multiple countries outside the United States, to advance our research and development efforts.
Current or future tariffs will result in increased research and development expenses, including with respect to increased costs associated with DS, DP, raw materials, laboratory equipment and research materials and components. In addition, such tariffs will increase our supply chain complexity and could also potentially disrupt our existing supply chain. Unlike consumer goods, pharmaceuticals face unique regulatory constraints that make rapid supply chain adjustments particularly difficult and costly. Trade restrictions affecting the import of materials necessary for clinical trials could result in delays to our development timelines. Increased development costs and extended development timelines could place us at a competitive disadvantage compared to companies operating in regions with more favorable trade relationships and could reduce investor confidence, negatively impacting our
75
Table of Contents
ability to secure additional financing on favorable terms or at all. In addition, as we advance toward commercialization in the future, tariffs and trade restrictions could hinder our ability to establish cost-effective production capabilities, negatively impacting our growth prospects.
The complexity of announced or future tariffs may also increase the risk that we or our customers or suppliers may be subject to civil or criminal enforcement actions in the United States or foreign jurisdictions related to compliance with trade regulations. Foreign governments may also adopt non-tariff measures, such as procurement preferences or informal disincentives to engage with, purchase from or invest in U.S. entities, which may limit our ability to compete internationally and attract non-U.S. investment, employees, customers and suppliers. Foreign governments may also take other retaliatory actions against U.S. entities, such as decreased intellectual property protection, increased enforcement actions, or delays in regulatory approvals, which may result in heightened international legal and operational risks. In addition, the United States and other governments have imposed and may continue to impose additional sanctions, such as trade restrictions or trade barriers, which could restrict us from doing business directly or indirectly in or with certain countries or parties and may impose additional costs and complexity to our business.
Trade disputes, tariffs, restrictions and other political tensions between the United States and other countries may also exacerbate unfavorable macroeconomic conditions including inflationary pressures, foreign exchange volatility, financial market instability, and economic recessions or downturns. The ultimate impact of current or future tariffs and trade restrictions remains uncertain and could materially and adversely affect our business, financial condition, results of operations, and prospects. While we actively monitor these risks, any prolonged economic downturn, escalation in trade tensions, or deterioration in international perception of U.S.-based companies could materially and adversely affect our business, ability to access the capital markets or other financing sources, results of operations, financial condition and prospects. In addition, tariffs and other trade developments have and may continue to heighten the risks related to the other risk factors described elsewhere in this registration statement.
We will incur significant increased costs as a result of operating as a public company, and our management will be required to devote substantial time to new compliance initiatives and corporate governance practices.
As a public company, and particularly after we are no longer an emerging growth company or smaller reporting company, we will incur significant legal, accounting and other expenses that we did not incur as a private company. The Sarbanes-Oxley Act, the Dodd-Frank Wall Street Reform and Consumer Protection Act, the listing requirements of Nasdaq and other applicable securities rules and regulations impose various requirements on public companies, including establishment and maintenance of effective disclosure and financial controls and corporate governance practices. Our management and other personnel will need to devote a substantial amount of time to these compliance initiatives. If these requirements divert the attention of our management and personnel from other business concerns, they could have a material adverse effect on our prospects, business, financial condition and results of operations. Moreover, we expect these rules and regulations to substantially increase our legal and financial compliance costs and to make some activities more time consuming and costly. For example, we expect that these rules and regulations may make it more difficult and more expensive for us to obtain director and officer liability insurance and we may be required to incur substantial costs to maintain sufficient coverage. We cannot predict or estimate the amount or timing of additional costs we may incur to respond to these requirements. The impact of these requirements could also make it more difficult for us to attract and retain qualified persons to serve on our board of directors, our board committees or as executive officers. The increased costs will decrease our net income or increase our net loss, and the increased costs may require us to reduce costs in other areas of our business.
Moreover, these rules and regulations are often subject to varying interpretations, in many cases due to their lack of specificity, and, as a result, their application in practice may evolve over time as new guidance is provided by regulatory and governing bodies. This could result in continuing uncertainty regarding compliance matters and higher costs necessitated by ongoing revisions to disclosure and governance practices.
We may be subject to securities litigation, which is expensive and could divert management attention.
The market price of our common stock is likely to be volatile. The stock market in general, and Nasdaq and biopharmaceutical companies in particular, have experienced extreme price and volume fluctuations that have often been unrelated or disproportionate to the operating performance of these companies. In the past, companies that
76
Table of Contents
have experienced volatility in the market price of their stock have been subject to securities class action litigation. We may be the target of this type of litigation in the future. Securities litigation against us could result in substantial costs, divert our management’s attention and resources from other business concerns and damage our reputation, which could seriously harm our business, financial condition, results of operations, and prospects. Additionally, the increase in the cost of directors’ and officers’ liability insurance may cause us to opt for lower overall policy limits or to forgo insurance that we may otherwise rely on to cover significant defense costs, settlements and damages awarded to plaintiffs.
We are an “emerging growth company” and a “smaller reporting company” and the reduced reporting requirements applicable to emerging growth companies or smaller reporting companies could make our common stock less attractive to investors.
We are an “emerging growth company” as defined in the JOBS Act. For as long as we continue to be an emerging growth company, we may take advantage of exemptions from various reporting requirements that are applicable to other public companies that are not emerging growth companies, including (i) not being required to comply with the auditor attestation requirements of Section 404 of the Sarbanes-Oxley Act, (ii) reduced disclosure obligations regarding executive compensation in this prospectus and our periodic reports and proxy statements and (iii) exemptions from the requirements of holding nonbinding advisory stockholder votes on executive compensation and stockholder approval of any golden parachute payments not approved previously. In addition, as an emerging growth company, we are only required to provide two years of audited financial statements and two years of selected financial data in this prospectus.
We could be an emerging growth company for up to five years following the completion of this offering, although circumstances could cause us to lose that status earlier, including if we are deemed to be a “large accelerated filer,” which occurs when the market value of our common stock that is held by non-affiliates equals or exceeds $700.0 million as of the prior June 30, or if we have total annual gross revenue of $1.235 billion or more during any fiscal year before that time, in which cases we would no longer be an emerging growth company as of the following December 31, or if we issue more than $1.0 billion in non-convertible debt during any three-year period before that time, in which case we would no longer be an emerging growth company immediately.
Under the JOBS Act, emerging growth companies can also delay adopting new or revised accounting standards until such time as those standards apply to private companies. We have elected to take advantage of the benefits of this extended transition period. Our financial statements may therefore not be comparable to those of companies that comply with such new or revised accounting standards. Until the date that we are no longer an “emerging growth company” or affirmatively and irrevocably opt out of the exemption provided by Section 7(a)(2)(B) of the Securities Act, upon issuance of a new or revised accounting standard that applies to our financial statements and that has a different effective date for public and private companies, we will disclose the date on which adoption is required for non-emerging growth companies and the date on which we will adopt the recently issued accounting standard.
We are also a “smaller reporting company” as defined in the Exchange Act. We may continue to be a smaller reporting company after this offering if either (i) the market value of our common stock held by non-affiliates is less than $250.0 million, measured as of the last business day of our most recently completed second quarter or (ii) our annual revenue is less than $100.0 million during the most recently completed fiscal year and the market value of our common stock held by non-affiliates is less than $700.0 million. We may continue to be a smaller reporting company even after we cease to be an emerging growth company, so we may continue to rely on exemptions from certain disclosure requirements that are available to smaller reporting companies. Specifically, as a smaller reporting company we may choose to present only the two most recent fiscal years of audited financial statements, we are not required to comply with the auditor attestation requirements of Section 404 of the Sarbanes-Oxley Act and, similar to emerging growth companies, smaller reporting companies have reduced disclosure obligations regarding executive compensation.
If we fail to establish and maintain proper and effective internal control over financial reporting in the future, our ability to produce accurate and timely financial statements could be impaired, which could harm our operating results, investors’ views of us and, as a result, the value of our common stock.
Pursuant to Section 404 of the Sarbanes-Oxley Act, we will be required to furnish a report by our management on our internal control over financial reporting within our Annual Report on Form 10-K. However, while we remain an
77
Table of Contents
emerging growth company, we will not be required to include an attestation report on internal control over financial reporting issued by our independent registered public accounting firm. Ensuring that we have adequate internal financial and accounting controls and procedures in place so that we can produce accurate financial statements on a timely basis is a costly and time-consuming effort that will need to be frequently evaluated. Our failure to maintain the effectiveness of our internal controls in accordance with the requirements of the Sarbanes-Oxley Act could have a material adverse effect on our business. If we identify one or more material weaknesses, it could result in an adverse reaction in the financial markets due to a loss of confidence in the reliability of our financial statements. In addition, if we are not able to continue to meet these requirements, we may not be able to remain listed on Nasdaq.
As we grow, we expect to hire additional personnel and may utilize external temporary resources to implement, document and modify policies and procedures to maintain effective internal controls. However, it is possible that we may identify deficiencies and weaknesses in our internal controls. If material weaknesses or deficiencies in our internal controls exist and go undetected or unremediated, our financial statements could contain material misstatements that, when discovered in the future, could cause us to fail to meet our future reporting obligations and cause the price of our common stock to decline.
Our disclosure controls and procedures may not prevent or detect all errors or acts of fraud.
Upon the completion of this offering, we will become subject to the periodic reporting requirements of the Exchange Act. We designed our disclosure controls and procedures to reasonably assure that information we must disclose in reports we file or submit under the Exchange Act is accumulated and communicated to management, and recorded, processed, summarized and reported within the time periods specified in the rules and forms of the SEC. We believe that any disclosure controls and procedures or internal controls and procedures, no matter how well-conceived and operated, can provide only reasonable, not absolute, assurance that the objectives of the control system are met.
These inherent limitations include the realities that judgments in decision-making can be faulty, and that breakdowns can occur because of simple error or mistake. For example, our directors or executive officers could inadvertently fail to disclose a new relationship or arrangement causing us to fail to make any related party transaction disclosures. Additionally, controls can be circumvented by the individual acts of some persons, by collusion of two or more people or by an unauthorized override of the controls. Accordingly, because of the inherent limitations in our control system, misstatements due to error or fraud may occur and not be detected. In addition, we do not have a formal risk management program for identifying and addressing risks to our business in other areas.
Our insurance policies may be inadequate, may not cover all of our potential liabilities and may potentially expose us to unrecoverable risks.
We do not carry insurance for all categories of risk that our business may encounter. Some of the policies we currently maintain include property, general liability, employee benefits liability, workers’ compensation, clinical trials/products liability, cyber liability, directors’ and officers’ and employment practices insurance. We do not know, however, if we will be able to maintain insurance with adequate levels of coverage. No assurance can be given that an insurance carrier will not seek to cancel or deny coverage after a claim has occurred. Any significant uninsured liability may require us to pay substantial amounts, which would adversely affect our financial position and results of operations. For example, although we maintain product liability insurance coverage that also covers our clinical trials, this insurance may not be adequate to cover all liabilities that we may incur, and we may be required to increase our product liability insurance coverage. We anticipate that we will need to increase our insurance coverage each time we commence a clinical trial and successfully commercialize any product candidate. Insurance availability, coverage terms and pricing continue to vary with market conditions. We endeavor to obtain appropriate insurance coverage for insurable risks that we identify. However, we may fail to correctly anticipate or quantify insurable risks, we may not be able to obtain appropriate insurance coverage and insurers may not respond as we intend to cover insurable events that may occur. Any significant uninsured liability may require us to pay substantial amounts, which would materially adversely affect our business, financial condition, results of operations, and growth.
78
Table of Contents
SPECIAL NOTE REGARDING FORWARD-LOOKING STATEMENTS
This prospectus contains forward-looking statements about us and our industry that involve substantial risks and uncertainties. All statements other than statements of historical fact contained in this prospectus, including statements regarding our plans, objectives, goals, strategies, future events, future revenues or performance, financing needs, plans, or intentions relating to our product candidates and development programs, and markets and business trends are forward-looking statements. We have based these forward-looking statements largely on our current expectations and projections. In some cases, you can identify forward-looking statements because they contain words such as “aim,” “anticipate,” “assume,” “believe,” “can,” “contemplate,” “continue,” “could,” “design,” “due,” “estimate,” “expect,” “goal,” “intend,” “may,” “might,” “objective,” “plan,” “positioned,” “potential,” “predict,” “project,” “seek,” “shall,” “should,” “target,” “will,” “would,” or the negative of these words or other similar terms or expressions.
These statements involve known and unknown risks, uncertainties, and other factors which may cause our actual results, performance, or achievements to be materially different from any future results, performance, or achievements expressed or implied by the forward-looking statements. Forward-looking statements include, but are not limited to, statements about:
| ∎ | the initiation, timing, progress, and results of our research and development programs, nonclinical studies, and clinical trials; |
| ∎ | our ability to develop and advance ipsoprubart and ELA822 and any future product candidates; |
| ∎ | the timing of and costs involved in obtaining and maintaining regulatory approval of ipsoprubart and ELA822 and any future product candidates that we may identify or develop; |
| ∎ | the beneficial characteristics, including potential safety, efficacy, and therapeutic effects of ipsoprubart and ELA822; |
| ∎ | our ability to efficiently and cost-effectively conduct our current and future nonclinical studies and clinical trials; |
| ∎ | our ability to obtain funding for our operations necessary to complete further development and, if approved, commercialization of ipsoprubart and ELA822; |
| ∎ | our ability to identify and develop ipsoprubart and ELA822 for the treatment of additional indications; |
| ∎ | the performance of our third-party service providers, including our suppliers and manufacturers; |
| ∎ | the rate and degree of market acceptance and clinical utility for ipsoprubart, ELA822 and any other product candidates we may develop; |
| ∎ | the effects of competition with respect to ipsoprubart and ELA822 or any other product candidates we may develop, as well as innovations by competitors in our industry; |
| ∎ | our estimates regarding the potential market opportunities and the number of patients for ipsoprubart, ELA822 and any future product candidates, if approved for commercial use; |
| ∎ | the implementation of our strategic plans for our business, ipsoprubart, ELA822, and any future product candidates we may develop; |
| ∎ | our intellectual property position, including the scope of protection we are able to establish, maintain, defend, and enforce for intellectual property rights covering ipsoprubart, ELA822, and any future product candidates we may develop; |
| ∎ | our ability to attract and retain key scientific or management personnel; |
| ∎ | regulatory and legal developments in the United States; |
| ∎ | our ability to attract and retain employees and collaborators with development, regulatory, and commercialization expertise; |
| ∎ | the accuracy of our estimates regarding future expenses, future revenue, capital requirements, and need for additional financing; |
| ∎ | the period over which we estimate our existing cash, cash equivalents, and marketable securities will be sufficient to fund our future operating expenses and capital expenditure requirements; |
79
Table of Contents
| ∎ | our expectations regarding the impact of global health pandemics, geopolitical conflicts, and economic uncertainty, including interest rate changes and inflation on our business and operations, including clinical trials, collaborators, CMOs, and employees; |
| ∎ | our expectations regarding the period during which we qualify as an emerging growth company under the JOBS Act; and |
| ∎ | our anticipated use of the proceeds from this offering. |
These forward-looking statements reflect our management’s beliefs and views with respect to future events and are based on estimates and assumptions as of the date of this prospectus and are subject to risks and uncertainties. In addition, statements that “we believe” and similar statements reflect our beliefs and opinions on the relevant subject. These statements are based upon information available to us as of the date of this prospectus, and while we believe such information forms a reasonable basis for such statements, such information may be limited or incomplete, and our statements should not be read to indicate that we have conducted an exhaustive inquiry into, or review of, all potentially available relevant information. These statements are inherently uncertain and investors are cautioned not to unduly rely upon these statements. We discuss many of the risks associated with the forward-looking statements in this prospectus in greater detail in the section titled “Risk Factors.” Moreover, we operate in a very competitive and rapidly changing environment. New risks emerge from time to time. It is not possible for our management to predict all risks, nor can we assess the impact of all factors on our business or the extent to which any factor, or combination of factors, may cause actual results to differ materially from those contained in any forward-looking statements we may make. Given these uncertainties, you should not place undue reliance on these forward-looking statements.
You should carefully read this prospectus and the documents that we have filed as exhibits to the registration statement, of which this prospectus is a part, completely and with the understanding that our actual future results may be materially different from what we expect. We qualify all of the forward-looking statements in this prospectus by these cautionary statements.
Except as required by law, we assume no obligation to update these forward-looking statements publicly, or to update the reasons actual results could differ materially from those anticipated in any forward-looking statements, whether as a result of new information, future events, or otherwise.
80
Table of Contents
MARKET, INDUSTRY, AND OTHER DATA
Certain market, industry, and competitive data included in this prospectus were obtained from our own internal estimates and research, as well as from publicly available information, reports of governmental agencies, and industry and academic publications, and research conducted by third parties. In some cases, we do not expressly refer to the sources from which this data is derived. Information that is based on estimates, forecasts, projections, market research, or similar methodologies is inherently subject to uncertainties and involves a number of assumptions and limitations; as a result, actual events or circumstances may differ materially from events and circumstances that are assumed in this information, and you are cautioned not to give undue weight to such information.
The industry in which we operate is subject to a high degree of uncertainty and risk due to a variety of factors, including those described in the section titled “Risk Factors.” These and other factors could cause results to differ materially from those expressed in estimates made by third parties or by us.
81
Table of Contents
We have never declared or paid any cash dividends on our capital stock and we do not currently intend to pay any cash dividends on our capital stock for the foreseeable future. We currently intend to retain all available funds and any future earnings to support operations and to finance the growth and development of our business. Any future determination to pay dividends will be made at the discretion of our board of directors, subject to applicable laws and will depend upon, among other factors, our results of operations, financial condition, contractual restrictions, and capital requirements.
82
Table of Contents
We estimate that we will receive net proceeds from this offering of approximately $296.6 million (or approximately $341.9 million if the underwriters exercise their option to purchase additional shares in full) assuming an initial public offering price of $15.00 per share of our common stock, which is the midpoint of the price range set forth on the cover page of this prospectus, after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us.
Each $1.00 increase (decrease) in the assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, would increase (decrease) the net proceeds to us from this offering by approximately $20.2 million, assuming the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same, and after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. We may also increase or decrease the number of shares we are offering. Similarly, each one million share increase (decrease) in the number of shares offered by us would increase (decrease) the net proceeds to us from this offering by approximately $14.0 million, assuming that the assumed initial offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, remains the same, and after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. We do not expect that a change in the initial price to the public or the number of shares by these amounts would have a material effect on the uses of the proceeds from this offering, although it may accelerate the time at which we will need to seek additional capital.
The principal purposes of this offering are to obtain additional capital to support our operations, to create a public market for our common stock, and to facilitate our future access to the public equity markets. We currently intend to use the net proceeds from this offering, together with our existing cash, cash equivalents, and marketable securities, as follows:
| ∎ | approximately $220.0 million to advance the development of ipsoprubart in sHLH through our ongoing Phase 2/3 global registrational trial topline results and to support BLA submission and commercial readiness activities; |
| ∎ | approximately $25.0 million to advance the development of ipsoprubart in T/NK cell malignancies through our ongoing Phase 1 clinical trial; |
| ∎ | approximately $50.0 million to advance the development of ELA822 through our ongoing Phase 1 clinical trial in healthy volunteers and initiation of a planned Phase 1/2 clinical trial in patients with T cell mediated immune disorders; and |
| ∎ | the remaining amounts for other research and development activities, as well as for working capital and other general corporate purposes. |
We may also use a portion of the net proceeds for strategic investments in complementary businesses, products, technologies, or assets. However, we do not have agreements or commitments to enter into any such acquisitions or investments at this time.
Based on our current operating plans, we estimate that our existing cash, cash equivalents, and marketable securities as of the date of this prospectus, together with the estimated net proceeds from this offering, will be sufficient to fund our projected operating expenses and capital expenditure requirements into 2029. We have based this estimate on assumptions that may prove to be wrong, and we could use our available capital resources sooner than we expect.
The expected net proceeds from this offering, together with our existing cash, cash equivalents, and marketable securities, will not be sufficient for us to fund our product candidates through clinical trials, regulatory approval, and commercialization, and we will need to raise substantial additional capital in order to do so.
We cannot predict with certainty all of the particular uses for the proceeds of this offering or the amounts that we will actually spend on the uses set forth above. Our management will have broad discretion over the use of the net proceeds from this offering. The amounts and timing of our actual use of the net proceeds will vary depending on
83
Table of Contents
numerous factors, including the progress, cost, and results of our preclinical and clinical development programs, our ability to obtain additional financing, and other factors described under “Risk Factors” in this prospectus, as well as the amount of cash used in our operations and any unforeseen cash needs. We may find it necessary or advisable to use the net proceeds for other purposes, and investors will be relying on the judgment of our management regarding the application of those net proceeds.
Pending the use of proceeds from this offering described above, we intend to invest the net proceeds in one or more of the following: money market funds, short- and intermediate-term, interest-bearing, investment-grade securities, certificates of deposit, or direct or guaranteed obligations of the U.S. government.
84
Table of Contents
The following table sets forth our cash, cash equivalents, and marketable securities and our capitalization as of June 30, 2026 on:
| ∎ | an actual basis; |
| ∎ | a pro forma basis, giving effect to (i) the automatic conversion of all outstanding shares of our redeemable convertible preferred stock as of June 30, 2026 into 39,404,292 shares of our common stock immediately prior to the closing of this offering and the related reclassification of the carrying value of our redeemable convertible preferred stock to permanent equity in connection with the closing of this offering and (ii) the filing and effectiveness of our amended and restated certificate of incorporation upon the closing of this offering; and |
| ∎ | a pro forma as adjusted basis, giving effect to (i) the pro forma adjustments set forth above and (ii) the issuance and sale of shares of our common stock in this offering at the assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. |
The pro forma and pro forma as adjusted information below is illustrative only, and our cash, cash equivalents, and marketable securities and capitalization following the closing of this offering will be adjusted based on the actual initial public offering price and other terms of this offering determined at pricing. You should read the information in this table together with our financial statements and the related notes included elsewhere in this prospectus and the section titled “Management’s Discussion and Analysis of Financial Condition and Results of Operations” and other financial information contained in this prospectus.
| (in thousands, except share and per share data) | AS OF JUNE 30, 2026 (UNAUDITED) |
|||||||||||
| ACTUAL | PRO FORMA | PRO FORMA AS ADJUSTED (1) |
||||||||||
| Cash, cash equivalents, and marketable securities |
$ | 97,725 | $ | 97,725 | $ | 394,325 | ||||||
|
|
|
|
|
|
|
|||||||
| Series A redeemable convertible preferred stock, $0.0001 par value; 22,601,626 shares authorized, 15,065,741 issued and outstanding, actual; no shares authorized, issued, or outstanding, pro forma and pro forma as adjusted |
34,650 | — | — | |||||||||
| Series B redeemable convertible preferred stock, $0.0001 par value; 9,309,492 shares authorized, 6,205,494 issued and outstanding, actual; no shares authorized, issued, or outstanding, pro forma and pro forma as adjusted |
83,836 | — | — | |||||||||
| Series C redeemable convertible preferred stock, $0.001 par value; 27,203,243 shares authorized, 18,133,057 issued and outstanding, actual; no shares authorized, issued, or outstanding, pro forma and pro forma as adjusted |
192,685 | — | — | |||||||||
| Stockholders’ deficit: |
||||||||||||
| Preferred stock, $0.0001 par value; no shares authorized, issued, or outstanding, actual; 10,000,000 shares authorized and no shares issued or outstanding, pro forma and pro forma as adjusted |
— | — | — | |||||||||
| Common stock, $0.0001 par value; 71,000,000 shares authorized, 29,329 shares issued and outstanding, actual; 900,000,000 shares authorized, pro forma and pro forma as adjusted; 39,433,621 shares issued and outstanding, pro forma; 61,100,288 shares issued and outstanding, pro forma as adjusted |
— | 4 | 6 | |||||||||
| Additional paid-in capital |
2,448 | 313,615 | 610,213 | |||||||||
| Accumulated deficit |
(221,890 | ) | (221,890 | ) | (221,890 | ) | ||||||
| Accumulated other comprehensive income (loss) |
(13 | ) | (13 | ) | (13 | ) | ||||||
|
|
|
|
|
|
|
|||||||
| Total stockholders’ equity (deficit) |
(219,455 | ) | 91,716 | 388,316 | ||||||||
|
|
|
|
|
|
|
|||||||
| Total capitalization |
$ | 91,716 | $ | 91,716 | $ | 388,316 | ||||||
|
|
|
|
|
|
|
|||||||
85
Table of Contents
| (1) | Each $1.00 increase (or decrease) in the assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, would increase (decrease) the pro forma as adjusted amount of each of our cash, cash equivalents, and marketable securities, additional paid-in capital, total stockholders’ equity, and total capitalization by approximately $20.2 million, assuming that the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same, and after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. We may also increase or decrease the number of shares we are offering. Each one million share increase (decrease) in the number of shares offered by us at the assumed initial public offering price per share of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, would increase (decrease) the pro forma as adjusted amount of each of our cash, cash equivalents, and marketable securities, additional paid-in capital, total stockholders’ equity, and total capitalization by approximately $14.0 million, after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us. |
The information in the table above excludes:
| ∎ | 5,701,943 shares of our common stock issuable upon the exercise of stock options outstanding under the 2022 Plan as of June 30, 2026, with a weighted-average exercise price of $4.92 per share; |
| ∎ | 259,921 shares of our common stock issuable upon the exercise of stock options outstanding granted under the 2022 Plan subsequent to June 30, 2026, with a weighted-average exercise price of $7.03 per share; |
| ∎ | 2,863,183 shares of our common stock issuable upon the exercise of the IPO Option Grants; |
| ∎ | 13,446,281 shares of our common stock reserved for future issuance under our 2026 Plan (inclusive of the shares issuable under the IPO Option Grants), which will become effective upon the execution and delivery of the underwriting agreement for this offering (which shares include 7,697,039 new shares plus the number of shares (not to exceed 5,749,242 shares) that remain available for the issuance of awards under the 2022 Plan, at the time the 2026 Plan becomes effective, as more fully described in the section titled “Executive and Director Compensation—Equity Benefit Plans”), as well as any automatic increases in the number of shares of our common stock reserved for future issuance under the 2026 Plan; and |
| ∎ | 611,682 shares of our common stock reserved for future issuance under our ESPP, which will become effective upon the execution and delivery of the underwriting agreement for this offering, as well as any annual automatic increases in the number of shares of our common stock reserved for future issuance under the ESPP. |
86
Table of Contents
If you invest in our common stock in this offering, your ownership interest will be immediately diluted to the extent of the difference between the initial public offering price per share of our common stock and the pro forma as adjusted net tangible book value per share of our common stock immediately after the closing of this offering.
As of June 30, 2026, we had a historical net tangible book value (deficit) of approximately $(221.4) million, or $(7,548.02) per share of our common stock based on 29,329 shares of our common stock outstanding as of such date. Our historical net tangible book value (deficit) per share represents the amount of our total tangible assets less our total liabilities and redeemable convertible preferred stock, divided by the total number of shares of our common stock outstanding as of June 30, 2026.
After giving effect to the automatic conversion of all outstanding shares of our redeemable convertible preferred stock into 39,404,292 shares of our common stock and the related reclassification of the carrying value of our redeemable convertible preferred stock to permanent equity in connection with the closing of this offering and assuming the occurrence of such conversion as of June 30, 2026, our pro forma net tangible book value as of June 30, 2026, would have been approximately $89.8 million, or approximately $2.28 per share of our common stock.
Net tangible book value dilution per share to new investors represents the difference between the amount per share paid by purchasers of shares of our common stock in this offering and the pro forma as adjusted net tangible book value per share of our common stock immediately after closing of this offering. After giving further effect to the sale of 21,666,667 shares of our common stock that we are offering at the assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, and after deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us, our pro forma as adjusted net tangible book value as of June 30, 2026 would have been $386.4 million, or approximately $6.32 per share of our common stock. This amount represents an immediate increase in pro forma net tangible book value of $4.04 per share to our existing stockholders and an immediate dilution in pro forma net tangible book value of approximately $8.68 per share to new investors participating in this offering.
Dilution per share to new investors is determined by subtracting pro forma as adjusted net tangible book value per share after this offering from the assumed initial public offering price per share paid by new investors. The following table illustrates this dilution:
| Assumed initial public offering price per share |
15.00 | |||||||
| Historical net tangible book value (deficit) per share as of June 30, 2026 |
$ | (7,548.02 | ) | |||||
| Pro forma increase in historical net tangible book value per share as of June 30, 2026 |
7,550.30 | |||||||
|
|
|
|||||||
| Pro forma net tangible book value per share as of June 30, 2026, before this offering |
2.28 | |||||||
| Increase in pro forma net tangible book value per share attributed to new investors purchasing shares in this offering |
4.04 | |||||||
|
|
|
|||||||
| Pro forma as adjusted net tangible book value per share immediately after this offering |
6.32 | |||||||
|
|
|
|||||||
| Dilution per share to new investors in this offering |
$ | 8.68 | ||||||
|
|
|
The dilution information discussed above is illustrative only and will change based on the actual initial public offering price and other terms of this offering determined at pricing. Each $1.00 increase (decrease) in the assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, would increase (decrease) the pro forma as adjusted net tangible book value per share after this offering by approximately $0.33, and dilution in pro forma net tangible book value per share to new investors by approximately $0.67, assuming that the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same and after deducting the estimated underwriting discounts and commissions and the estimated offering expenses payable by us.
87
Table of Contents
We may also increase or decrease the number of shares we are offering. Each increase of one million shares in the number of shares offered by us, as set forth on the cover page of this prospectus, would increase our pro forma as adjusted net tangible book value per share after this offering by approximately $0.13 and decrease the dilution to investors participating in this offering by approximately $0.13 per share, assuming that the assumed initial public offering price remains the same, and after deducting the estimated underwriting discounts and commissions and the estimated offering expenses payable by us. Similarly, each decrease of one million shares in the number of shares offered by us, as set forth on the cover page of this prospectus, would decrease the pro forma as adjusted net tangible book value per share after this offering by approximately $0.12 and increase the dilution to investors participating in this offering by approximately $0.12 per share, assuming the assumed initial public offering price of $15.00 per share remains the same, and after deducting the estimated underwriting discounts and commissions and the estimated offering expenses payable by us.
If the underwriters exercise their option to purchase up to 3,250,000 additional shares of our common stock in full in this offering, the pro forma as adjusted net tangible book value after the offering would be $6.71 per share, the increase in pro forma as adjusted net tangible book value per share to existing stockholders would be $4.43 per share and the dilution per share to new investors would be $8.29 per share, in each case assuming an initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, remains the same and after deducting the estimated underwriting discounts and commissions payable by us.
To the extent that outstanding options with an exercise price per share that is less than the pro forma as adjusted net tangible book value per share are exercised, new investors will experience further dilution. In addition, we may choose to raise additional capital due to market conditions or strategic considerations even if we believe we have sufficient funds for our current or future operating plans. To the extent that we raise additional capital through the sale of equity or convertible debt securities, the issuance of these securities could result in further dilution to our stockholders.
The following table summarizes, on the pro forma as adjusted basis described above, as of June 30, 2026, the differences between the number of shares of our common stock purchased from us by our existing stockholders and common stock by new investors purchasing shares in this offering, the total consideration paid to us in cash and the weighted-average price per share paid by existing stockholders for shares of our common stock issued prior to this offering and the price to be paid by new investors for shares of our common stock in this offering. The calculation below is assuming an initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover page of this prospectus, before deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us.
| SHARES PURCHASED | TOTAL CONSIDERATION | WEIGHTED- AVERAGE PRICE PER |
||||||||||||||||||
| NUMBER | PERCENT | AMOUNT (IN THOUSANDS) |
PERCENT | SHARE | ||||||||||||||||
| Existing stockholders |
39,433,621 | 64.5 | % | $ | 301,522 | 48.1 | % | $ | 7.65 | |||||||||||
| New investors |
21,666,667 | 35.5 | % | $ | 325,000 | 51.9 | 15.00 | |||||||||||||
| Total |
61,100,288 | 100.0 | % | $ | 626,522 | 100.0 | % | $ | 10.25 | |||||||||||
|
|
|
|
|
|
|
|||||||||||||||
Each $1.00 increase in the assumed initial public offering price of $15.00 per share would increase total consideration paid by new investors, total consideration paid by all stockholders and the weighted-average price per share paid by all stockholders by $21.7 million, $21.7 million and $0.35, respectively, while each $1.00 decrease in the assumed initial public offering price of $15.00 per share, would decrease total consideration paid by new investors, total consideration paid by all stockholders and the weighted-average price per share paid by all stockholders by $21.7 million, $21.7 million and $0.35, respectively, and assuming the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same and before deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us.
88
Table of Contents
Similarly, each one million share increase in the number of shares offered by us, as set forth on the cover page of this prospectus, would increase the total consideration paid by investors participating in this offering, total consideration paid by all stockholders, and the weighted-average price per share paid by all stockholders by approximately $15.0 million, $15.0 million, and $0.08, respectively, while each one million share decrease in the number of shares offered by us, as set forth on the cover page of this prospectus, would decrease the total consideration paid by investors participating in this offering, total consideration paid by all stockholders, and the weighted-average price per share paid by all stockholders by approximately $15.0 million, $15.0 million, and $0.08, respectively, assuming the assumed initial public offering price of $15.00 per share remains the same, and before deducting the estimated underwriting discounts and commissions and estimated offering expenses payable by us.
The table above assumes no exercise of the underwriters’ option to purchase additional shares of our common stock in this offering. If the underwriters were to fully exercise their option to purchase additional shares of our common stock from us, the percentage of our common stock held by existing stockholders after this offering would be reduced to 61.3% of the total number of shares of our common stock outstanding after this offering, and the percentage of our common stock held by new investors would be increased to 38.7% of the total number of shares of our common stock outstanding after this offering.
Except as otherwise indicated, the discussion and the tables above assume no exercise of the underwriters’ option to purchase additional shares of our common stock and excludes:
| ∎ | 5,701,943 shares of our common stock issuable upon the exercise of outstanding stock options as of June 30, 2026, with a weighted-average exercise price of $4.92 per share; |
| ∎ | 259,921 shares of our common stock issuable upon exercise of outstanding stock options granted subsequent to June 30, 2026, with a weighted-average exercise price of $7.03 per share; |
| ∎ | 2,863,183 shares of our common stock issuable upon the exercise of the IPO Option Grants; |
| ∎ | 13,446,281 shares of our common stock reserved for future issuance under our 2026 Plan (inclusive of the shares issuable under the IPO Option Grants), which will become effective upon the execution and delivery of the underwriting agreement for this offering (which shares include 7,697,039 new shares plus the number of shares (not to exceed 5,749,242 shares) that remain available for the issuance of awards under the 2022 Plan at the time the 2026 Plan becomes effective, as more fully described in the section titled “Executive and Director Compensation—Equity Benefit Plans”), as well as any automatic increases in the number of shares of our common stock reserved for future issuance under the 2026 Plan; and |
| ∎ | 611,682 shares of our common stock reserved for future issuance under our ESPP, as well as any annual automatic increases in the number of shares of our common stock reserved for future issuance under the ESPP, which will become effective upon the execution and delivery of the underwriting agreement for this offering. |
We may choose to raise additional capital through the sale of equity or convertible debt securities due to market conditions or strategic considerations even if we believe we have sufficient funds for our current or future operating plans. To the extent that stock options are exercised or we issue additional shares of our common stock or other equity or convertible debt securities in the future, there will be further dilution to investors participating in this offering.
89
Table of Contents
MANAGEMENT’S DISCUSSION AND ANALYSIS OF FINANCIAL CONDITION AND RESULTS OF OPERATIONS
The following discussion and analysis of our financial condition and results of operations should be read together with our financial statements and related notes and other financial information included elsewhere in this prospectus. This discussion and analysis and other parts of this prospectus contain forward-looking statements based upon current beliefs, plans and expectations related to future events and our future financial performance that involve risks, uncertainties and assumptions, such as statements regarding our intentions, plans, objectives, expectations and projections. Our actual results and the timing of certain events could differ materially from those discussed in the forward-looking statements as a result of many factors, including those discussed in the section titled “Risk Factors” and elsewhere in this prospectus. You should carefully read the section titled “Risk Factors” to gain an understanding of the important factors that could cause actual results to differ materially from our forward-looking statements. Please also see the section titled “Special Note Regarding Forward-Looking Statements.”
Overview
We are a late clinical-stage biopharmaceutical company focused on pioneering a new class of precision medicines for the treatment of immune-mediated diseases and cancer. Our novel approach targets signal regulatory proteins (SIRP), a family of cell surface receptors whose expression is restricted to specific immune cell populations, and increases upon activation to enable selective depletion of disease-driving cells while preserving normal immune function. By replacing broad immunosuppression with selective elimination of principal cells that drive disease, we believe our approach can do for immune-mediated diseases what precision oncology has done for cancer, transforming the treatment paradigm for patients. Our lead product candidate is ipsoprubart, a novel pan-SIRP monoclonal antibody designed to selectively deplete pathological myeloid cells and T cells via binding to SIRPa/ß1/g, which is currently in a global registrational program for patients with secondary hemophagocytic lymphohistiocytosis (sHLH). We are conducting SURPASS, our global Phase 2/3 registrational trial in newly diagnosed, treatment-naïve sHLH patients, as well as COMPASS, our natural history study designed to provide an external control comparator for SURPASS. We expect to complete enrollment in our registrational program in the second half of 2027. We are also advancing ELA822, a novel SIRPg-specific monoclonal antibody designed to selectively deplete activated T cells, with potential applications across chronic T cell-mediated immune and inflammatory diseases. In August 2026, we obtained regulatory clearance and initiated a Phase 1 clinical trial of ELA822 in healthy volunteers in Europe, with data expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027, subject to regulatory approval.
We commenced operations in 2018 and since our inception, we have devoted substantially all of our resources to organizing and staffing our company, business planning, raising capital, discovering our product candidates, establishing and maintaining our intellectual property portfolio, conducting research, nonclinical studies and clinical trials, establishing arrangements with third parties for the manufacture of our product candidates and related raw materials, and providing general and administrative support for these operations. To date, we have financed our operations primarily through the issuance and sale of shares of our redeemable convertible preferred stock and issuance of convertible promissory notes. As of June 30, 2026, issuances and sales of our redeemable convertible preferred stock have resulted in aggregate gross proceeds of $301.4 million. As of June 30, 2026, we had cash, cash equivalents and marketable securities of $97.7 million, which includes proceeds received from the recent second closing of our Series C redeemable convertible preferred stock financing, where we issued 4,918,477 shares of our Series C redeemable convertible preferred stock at a purchase price of $10.15 per share, resulting in gross proceeds of $49.9 million. Without giving effect to the anticipated net proceeds from this offering, based on our current operating plan, we estimated that our existing cash, cash equivalents and marketable securities as of June 30, 2026 may not have been sufficient to fund our projected operating expenses and capital expenditures for the next 12 months and, therefore, have concluded that circumstances exist that raise substantial doubt about our ability to continue as a going concern. However, based on our current operating plan, we estimate that our existing cash, cash equivalents and marketable securities as of the date of this prospectus, together with the estimated net proceeds from this offering, will be sufficient to fund our projected operating expenses and capital expenditures into 2029. This estimate is a forward-looking statement that involves risks and uncertainties, and actual results could vary materially.
90
Table of Contents
Since our inception, we have incurred significant net losses and negative cash flows from operations. We incurred net losses of $48.9 million and $25.4 million for the six months ended June 30, 2026 and 2025, respectively. We used $38.9 million and $24.7 million of cash in operating activities for the six months ended June 30, 2026 and 2025, respectively. As of June 30, 2026, we had an accumulated deficit of $221.9 million. To date, we have funded our operations primarily from the sales and issuances of our redeemable convertible preferred stock.
We expect our expenses and operating losses will increase substantially for the foreseeable future as we:
| ∎ | advance our product candidates through our ongoing nonclinical and clinical development; |
| ∎ | continue our research and development efforts and expand our pipeline of product candidates; |
| ∎ | maintain, expand and protect our intellectual property portfolio; |
| ∎ | collaborate with our contract manufacturing organizations to develop manufacturing processes and methods and establish manufacturing capacity to supply clinical trials in our pipeline and eventually for commercialization; |
| ∎ | seek regulatory approval for any potential future product candidates for which we successfully complete clinical trials; |
| ∎ | attract, hire and retain additional research and development, clinical, commercial and operational personnel; |
| ∎ | potentially experience any delays, challenges or other issues associated with the clinical development of our product candidates, including with respect to our regulatory strategies; |
| ∎ | make milestone, royalty or other payments under current, and any future, license or collaboration agreements; |
| ∎ | potentially seek to identify, acquire or in-license new technologies or product candidates; |
| ∎ | utilize third parties to manufacture our product candidates and related raw materials; |
| ∎ | implement operational, financial and management information systems; |
| ∎ | ultimately establish a sales, marketing and distribution infrastructure to commercialize any product candidate for which we may obtain marketing approval; and |
| ∎ | incur additional costs associated with being a public company, including audit, legal, regulatory and tax-related services associated with maintaining compliance with an exchange listing and U.S. Securities and Exchange Commission (SEC) requirements, director and officer insurance premiums and investor relations costs. |
If we obtain regulatory approval for any of our product candidates, we expect to incur significant commercialization expenses related to product sales, marketing, manufacturing, and distribution. Our net losses may fluctuate significantly from period to period, depending on the timing of and level of expense related to our clinical trials and nonclinical studies and our other research and development activities and capital expenditures and the timing and amount of any milestone or royalty payments due under our existing or future license or collaboration agreements. Cash used to fund operating expenses is impacted by the timing of payments for these expenses, as reflected in the change in our accounts payable and accrued liabilities.
We do not have any products approved for sale and have not generated any revenue to date. We do not expect to generate revenue from any product candidates until we successfully complete development and obtain regulatory approval for one or more of such product candidates, which we expect will take a number of years and may never occur. If we obtain regulatory approval for any of our product candidates, we expect to incur significant commercialization expenses related to product sales, marketing, manufacturing and distribution. As a result, we will need substantial additional funding to support our operating activities as we advance our product candidates through clinical development, seek regulatory approval and prepare for commercialization if any of our product candidates are approved. Until such time when we can generate significant revenue from sales of our product candidates, if ever, we expect to fund our operations through public or private equity offerings, debt financings, or potentially other capital sources, such as collaboration or licensing arrangements with third parties or other strategic transactions. However, we may not be able to raise additional funds or enter into such other arrangements when needed or on favorable terms, or at all. To the extent that we raise additional capital through the sale of equity or convertible debt
91
Table of Contents
securities, the ownership interest of our stockholders will be or could be diluted, and the terms of these securities may include liquidation or other preferences that adversely affect the rights of our common stockholders. Debt financing and equity financing, if available, may involve agreements that include covenants limiting or restricting our ability to take specific actions, such as incurring additional debt, making capital expenditures or declaring dividends. If we raise additional funds through future collaboration or licensing arrangements with third parties or other strategic transactions, we may have to relinquish rights to our intellectual property, future revenue streams, research programs or product candidates, or grant licenses on terms that may not be favorable to us and/or may reduce the value of our common stock. If we are unable to obtain adequate funding as and when needed, or on attractive terms, we could be required to significantly delay, reduce, or eliminate some or all of our research and development activities, product portfolio expansion or commercialization efforts, out-license intellectual property rights to our product candidates, sell unsecured assets, or scale back or terminate our pursuit of new strategic arrangements and transactions, or a combination of the above, any of which may have a material adverse effect on our business, financial condition, results of operations, and prospects.
We do not own or operate and currently have no plans to establish any manufacturing facilities. We rely and expect to continue to rely on third parties for the manufacture of our product candidates for nonclinical and clinical testing, as well as for commercial manufacture if our other product candidates obtain marketing approval. We are working with our current manufacturers to ensure that we will be able to scale up our manufacturing capabilities to support our clinical plans. In addition, we rely on third parties to package, label, store, and distribute our product candidates, and we intend to rely on third parties for our commercial products if marketing approval is obtained. We believe that this strategy allows us to maintain a more efficient infrastructure by eliminating the need for us to invest in our own manufacturing facilities, equipment, and personnel while also enabling us to focus our expertise and resources on the discovery and development of our product candidates.
Components of Our Results of Operations
Operating Expenses
Our operating expenses consist of (i) research and development expenses and (ii) general and administrative expenses.
Research and Development
Research and development expenses consist primarily of costs incurred in connection with the development of our nonclinical and clinical product candidates. Research and development expenses consist of:
Direct program costs:
| ∎ | expenses incurred under agreements with consultants and third-party contract organizations that conduct research and development activities on our behalf; |
| ∎ | costs related to the production of nonclinical and clinical materials related to our current or future product candidates paid to contract manufacturers; |
| ∎ | laboratory, testing and other expenses related to the execution of nonclinical studies and clinical trials; and |
| ∎ | third-party license fees, including any milestone-based payments. |
Unallocated costs:
| ∎ | personnel-related costs, such as salaries, benefits and stock-based compensation for employees engaged in research and development activities; and |
| ∎ | facility-related, overhead, and other costs not directly tied to a program, including rent, facility-related overhead, depreciation expense, expenses related to regulatory compliance and requirements, laboratory operations, information technology-related expenses, and other supplies and services. |
We expense all research and development costs in the period in which such costs are incurred. Costs for certain research and development activities are recognized based on evaluating the progress to completion of specific tasks using information and data provided to us by our vendors and third-party service providers. A significant portion of our research and development expenses have been direct external costs, which we track by program. However, we do
92
Table of Contents
not track our internal research and development expenses on a program specific basis because these costs are deployed across multiple projects and, as such, are not separately classified.
Non-refundable advance payments for goods and services used over time for research and development are capitalized and recognized as goods are delivered or as the related services are performed. Certain prepaid amounts are classified as long-term based on contractual terms and conditions. In-licensing fees and other costs to acquire technologies used in research and development that have not yet received regulatory approval and that are not expected to have an alternative future use are expensed when incurred.
Research and development activities are central to our business model, and the successful development of our product candidates is highly uncertain. As of the date of this prospectus, we cannot reasonably estimate or know the nature, timing and estimated costs of the efforts that will be necessary to complete the development of any of our product candidates. There are numerous factors associated with the approval and successful commercialization of any of our product candidates, including future trial design and various regulatory requirements, many of which cannot be determined with accuracy at this time based on our stage of development. Potential future changes to regulatory factors beyond our control may impact our development programs. Product candidates in later stages of clinical development generally have higher development costs than those in earlier stages of clinical development, primarily due to the increased size and duration of later-stage clinical trials. In addition, we cannot predict which product candidates may be subject to future collaborations, when such arrangements will be secured, if at all, and to what degree such arrangements would affect our development plans and capital requirements. We expect that our research and development expenses will increase substantially for the foreseeable future as we continue to invest in research and development activities related to developing our product candidates, as our product candidates advance into later and through various stages of development, as we seek regulatory approvals for any product candidates that successfully complete clinical trials, as we expand our product pipeline, as we maintain, expand, protect and enforce our intellectual property portfolio, and as we incur expenses associated with hiring additional personnel to support our research and development efforts.
The process of conducting the necessary clinical research to obtain regulatory approval is costly and time-consuming, and the successful development of our product candidates is highly uncertain. Our research and development expenses may vary significantly based on factors such as:
| ∎ | the number and scope of nonclinical and IND- or similar-enabling studies to enable the authorization of a similar regulatory submission outside the United States; |
| ∎ | the phases of development of our product candidates; |
| ∎ | the progress and results of our research and development activities; |
| ∎ | the number of trials required for regulatory approval; |
| ∎ | the number of sites included in each trial; |
| ∎ | the length of time required to initiate clinical trials and enroll eligible participants; |
| ∎ | the number of participants that participate in each trial; |
| ∎ | the drop-out and discontinuation rate of participants; |
| ∎ | potential additional safety monitoring requested by regulatory authorities; |
| ∎ | the duration of subject participation in the trials and follow-up; |
| ∎ | the cost and timing of manufacturing of our product candidates; |
| ∎ | the timing of licensing milestone payments related to development, regulatory and commercial events; |
| ∎ | the receipt of regulatory approvals from applicable regulatory authorities; |
| ∎ | the hiring and retention of research and development personnel; |
| ∎ | the degree to which we obtain, maintain, defend, and enforce our intellectual property rights; and |
| ∎ | the extent to which we establish collaboration, licensing or similar arrangements. |
A change in the outcome of any of these variables with respect to the development of any of our product candidates could significantly change the costs and timing associated with the development of that product candidate.
93
Table of Contents
General and Administrative
General and administrative expenses support our key business functions as we grow and mature as a company. General and administrative expenses consist of:
| ∎ | personnel-related costs, such as salaries, benefits and stock-based compensation for employees engaged in general and administrative activities; |
| ∎ | third-party service providers costs, including fees paid for legal, audit, accounting, consulting and other professional services; and |
| ∎ | facility-related, overhead, and other costs, including rent, facility-related overhead, depreciation expense, insurance costs, information technology-related expenses, and other supplies and services. |
We expect that our general and administrative expenses will increase substantially for the foreseeable future to support our expanding headcount and operations as we advance our product candidates through clinical development and commence our pre-commercial planning activities. Following the closing of this offering, we also expect that our costs will increase as a result of operating as a public company related to legal, audit, accounting, regulatory and tax-related services associated with maintaining compliance with exchange listing and SEC requirements, director and officer insurance costs, investor and public relations costs, and other expenses that we did not incur as a private company.
Other Income (Expense), Net
Our other income, expense, net consists of (i) interest income, (ii) interest expense and (iii) other income (expense).
Interest Income
Interest income consists primarily of interest from our marketable securities and money market accounts.
Interest Expense
Interest expense consists of the difference between the proceeds received from convertible promissory notes issued in March and June 2025 and the fair value of the convertible promissory notes, which was due to accrued interest on the convertible promissory notes as of the conversion date in October 2025.
Other Income (Expense)
Other income (expense) consists of foreign currency gains and losses.
Other Comprehensive Income
Other comprehensive income includes unrealized gains and losses on marketable securities.
94
Table of Contents
Results of Operations
Comparison of the Six Months Ended June 30, 2025 and 2026
The following table summarizes our statements of operations for the periods presented (in thousands, except percentages):
| SIX MONTHS ENDED JUNE 30, |
CHANGE | |||||||||||||||
| 2025 | 2026 | $ | % | |||||||||||||
| Operating expenses: |
||||||||||||||||
| Research and development |
$ | 18,792 | 41,637 | $ | 22,845 | 122% | ||||||||||
| General and administrative |
5,921 | 8,593 | 2,672 | 45% | ||||||||||||
| General and administrative – related party |
656 | — | (656 | ) | (100%) | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total operating expenses |
25,369 | 50,230 | 24,861 | 98% | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Loss from operations |
(25,369 | ) | (50,230 | ) | (24,861 | ) | 98% | |||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Other income (expense), net: |
||||||||||||||||
| Interest income |
337 | 1,320 | 983 | 292% | ||||||||||||
| Interest expense – related party |
(327 | ) | — | 327 | (100%) | |||||||||||
| Other income (expense) |
(2 | ) | (2 | ) | — | 0% | ||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total other income (expense), net |
8 | 1,318 | 1,310 | 16,375% | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Net loss |
$ | (25,361 | ) | (48,912 | ) | $ | (23,551 | ) | 93% | |||||||
|
|
|
|
|
|
|
|
|
|||||||||
Operating Expenses
Research and Development Expenses
The following table summarizes our research and development expenses for the periods presented (in thousands, except percentages):
| SIX MONTHS ENDED JUNE 30, |
CHANGE | |||||||||||||||
| 2025 | 2026 | $ | % | |||||||||||||
| Direct program costs: |
||||||||||||||||
| Ipsoprubart |
$ | 8,154 | $ | 26,124 | $ | 17,970 | 220% | |||||||||
| Other programs |
5,385 | 7,546 | 2,161 | 40% | ||||||||||||
| Unallocated costs: |
||||||||||||||||
| Personnel-related |
4,251 | 6,581 | 2,330 | 55% | ||||||||||||
| Facilities, overhead and other |
1,002 | 1,386 | 384 | 38% | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total research and development |
18,792 | 41,637 | 22,845 | 122% | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
Research and development expenses increased from $18.8 million for the six months ended June 30, 2025 to $41.6 million for the six months ended June 30, 2026. This increase was primarily attributable to the following:
| ∎ | an $18.0 million increase in expenses related to our ipsoprubart program, primarily driven by higher CRO costs, CMO costs, personnel-related expenses and consulting fees associated with the advancement of ipsoprubart into its Phase 2/3 clinical trial, including site initiation and patient enrollment activities that began in mid-2025 and have continued through 2026; |
| ∎ | a $2.2 million increase in expenses related to our other development programs, primarily due to higher CMO costs, material costs, and consulting fees incurred in programs unrelated to ipsoprubart; |
| ∎ | a $2.3 million increase in personnel-related expenses, primarily attributable to increased headcount to support the advancement of our research and development activities; and |
95
Table of Contents
| ∎ | a $0.4 million increase in other research and development expenses, primarily driven by higher rent expense and costs associated with increased headcount. |
General and Administrative Expenses
The following table summarizes our general and administrative expenses for the periods presented (in thousands, except percentages):
| SIX MONTHS ENDED JUNE 30, |
CHANGE | |||||||||||||||
| 2025 | 2026 | $ | % | |||||||||||||
| Personnel-related |
$ | 1,577 | 3,353 | $ | 1,776 | 113 | % | |||||||||
| Third-party service providers |
4,689 | 4,206 | (483 | ) | (10 | %) | ||||||||||
| Facilities, overhead and other |
311 | 1,034 | 723 | 232 | % | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total general and administrative |
6,577 | 8,593 | 2,016 | 31 | % | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
General and administrative expenses increased from $6.6 million for the six months ended June 30, 2025 to $8.6 million for the six months ended June 30, 2026. This increase was primarily attributable to the following:
| ∎ | a $1.8 million increase in personnel-related expenses, primarily driven by increased general and administrative headcount to support our growth; and |
| ∎ | a $0.7 million increase in other general and administrative expenses, primarily driven by higher rent expense and costs associated with increased headcount; partially offset by |
| ∎ | a $0.5 million decrease in fees for third-party service providers, primarily due to higher legal and accounting expenses incurred in 2025 in relation to financing activities. |
Other Income (Expense), Net
Interest Income
Interest income increased from $0.3 million for the six months ended June 30, 2025 to $1.3 million for the six months ended June 30, 2026. This increase was primarily due to higher average balances of marketable securities and higher interest rates and returns.
Interest Expense
Interest expense decreased from $0.3 million for the six months ended June 30, 2025 to zero for the six months ended June 30, 2026. The decrease was due to the difference between the proceeds received from convertible promissory notes issued in March and June 2025 and the fair value of the notes as of June 30, 2025. There was no interest expense during the six months ended June 30, 2026 on these convertible promissory notes, as the convertible promissory notes were fully converted to our Series C redeemable convertible preferred stock in October 2025.
Other Income (Expense)
Other expense, net was less than $0.1 million for the six months ended June 30, 2026 and 2025 and related primarily to foreign currency gains and losses.
Liquidity and Capital Resources
Sources of Liquidity
Since our inception, we have incurred significant net losses and negative cash flows in each period and on an aggregate basis. We have incurred significant net operating losses from operations since inception and had an accumulated deficit of $221.9 million as of June 30, 2026. Additionally, we anticipate that our operating losses and negative operating cash flows will increase for the foreseeable future as we continue to expand our research and development programs.
Since our inception, we have devoted substantially all of our resources to organizing and staffing our company, business planning, identifying product candidates, establishing our intellectual property portfolio, conducting
96
Table of Contents
research and nonclinical studies, including IND-enabling studies, initiating and conducting clinical trials, establishing arrangements with third parties for the manufacture of our product candidates, and providing general and administrative support for these operations.
To date, we have financed our operations primarily through the issuance and sale of shares of our redeemable convertible preferred stock and issuance of convertible promissory notes. As of June 30, 2026, issuances and sales of our redeemable convertible preferred stock have resulted in aggregate gross proceeds of $301.4 million.
On June 18, 2026, we completed the second closing of our Series C redeemable convertible preferred stock financing and issued 4,918,477 shares of our Series C redeemable convertible preferred stock at a purchase price of $10.15 per share, resulting in gross proceeds of $49.9 million.
Funding Requirements
As of June 30, 2026, we had cash, cash equivalents and marketable securities of $97.7 million.
We have incurred recurring losses and negative cash flows from operations and expect to continue to incur losses for the foreseeable future. Without giving effect to the anticipated net proceeds from this offering, we estimate that our existing cash, cash equivalents and marketable securities as of June 30, 2026 will not be sufficient to fund our operations for the next 12 months and, therefore, have concluded that circumstances exist that raise substantial doubt about our ability to continue as a going concern.
Our principal uses of cash in recent periods have been funding our research and development activities, including personnel-related costs, and general and administrative expenses. Our future capital requirements will depend on many factors, including, but not limited to:
| ∎ | the scope, progress, results and costs of researching, developing and manufacturing our product candidates or any future product candidates, and conducting nonclinical studies; |
| ∎ | the number and characteristics of any additional product candidates we develop or acquire; |
| ∎ | the timing of, and the costs involved in, obtaining regulatory approvals or clearances for our product candidates or any future product candidates; |
| ∎ | the expenses needed to attract and retain skilled personnel; |
| ∎ | the cost of any future product candidates and any products we successfully commercialize; |
| ∎ | the timing, receipt and amount of sales of any future approved or cleared products, if any; and |
| ∎ | our ability to establish and maintain strategic collaborations, licensing or other arrangements and the financial terms of any such agreements that we may enter into, including the timing and amount of any future milestone, royalty or other payments due under any such agreement. |
We may not be able to raise additional funds or enter into such other arrangements when needed or on favorable terms, or at all. To the extent that we raise additional capital through the sale of equity or convertible debt securities, the ownership interest of our stockholders will be or could be diluted, and the terms of these securities may include liquidation or other preferences that adversely affect the rights of our common stockholders. Debt financing and equity financing, if available, may involve agreements that include covenants limiting or restricting our ability to take specific actions, such as incurring additional debt, making capital expenditures or declaring dividends. If we raise additional funds through future collaboration or licensing arrangements with third parties or other strategic transactions, we may have to relinquish rights to our intellectual property, future revenue streams, research programs or product candidates, or grant licenses on terms that may not be favorable to us and/or may reduce the value of our common stock. If we are unable to obtain adequate funding as and when needed, or on attractive terms, we could be required to significantly delay, reduce, or eliminate some or all of our research and development activities, product portfolio expansion or commercialization efforts, out-license intellectual property rights to our product candidates, sell unsecured assets, or scale back or terminate our pursuit of new strategic arrangements and transactions, or a combination of the above, any of which may have a material adverse effect on our business, results of operations, and financial condition.
97
Table of Contents
Cash Flow Summary
The following table summarizes our cash flows for the periods presented (in thousands):
| SIX MONTHS ENDED JUNE 30, |
CHANGE | |||||||||||||||
| 2025 | 2026 | $ | % | |||||||||||||
| Net cash provided by (used in): |
||||||||||||||||
| Operating activities |
$ | (24,688 | ) | $ | (38,879 | ) | $ | (14,191 | ) | 57 | % | |||||
| Investing activities |
881 | 22,771 | 21,890 | 2,485 | % | |||||||||||
| Financing activities |
40,000 | 49,607 | 9,607 | 24 | % | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Net increase in cash and cash equivalents |
$ | 16,193 | $ | 33,499 | $ | 17,306 | 107 | % | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
Operating Activities
Net cash used in operating activities for the six months ended June 30, 2026 of $38.9 million was primarily due to our net loss of $48.9 million and $0.5 million of net amortization of premiums and discounts of marketable securities, partially offset by a $7.5 million net change in operating assets and liabilities, $2.6 million non-cash stock-based compensation expense, $0.3 million non-cash lease expense, and $0.1 million depreciation expense.
Net cash used in operating activities for the six months ended June 30, 2025 of $24.7 million was primarily due to our net loss of $25.4 million and $0.5 million of net decrease in operating assets and liabilities, partially offset by $0.9 million non-cash stock-based compensation expense and $0.3 million non-cash interest expense related to convertible promissory notes.
Investing Activities
Net cash provided by investing activities was $22.8 million for the six months ended June 30, 2026, which consisted primarily of $37.4 million of proceeds from the maturity of marketable securities, partially offset by $14.6 million in purchases of marketable securities.
Net cash provided by investing activities was $0.9 million for the six months ended June 30, 2025, which consisted primarily of $1.0 million of proceeds from marketable securities, offset by $0.1 million in purchases of equipment.
Financing Activities
Net cash provided by financing activities was $49.6 million for the six months ended June 30, 2026, which was attributable to the net proceeds from the issuance and sale of shares of our Series C redeemable convertible preferred stock in the Second Tranche Closing.
Net cash provided by financing activities was $40.0 million for the six months ended June 30, 2025, resulting from the issuance of our convertible promissory notes, which were fully converted during the year ended December 31, 2025 into shares of our Series C redeemable convertible preferred stock.
98
Table of Contents
Comparison of the Years Ended December 31, 2024 and 2025
The following table summarizes our statements of operations for the periods indicated (in thousands, except percentages):
| YEAR ENDED DECEMBER 31, |
CHANGE | |||||||||||||||
| 2024 | 2025 | $ | % | |||||||||||||
| Operating expenses: |
||||||||||||||||
| Research and development |
$ | 19,267 | $ | 50,387 | $ | 31,120 | 162% | |||||||||
| General and administrative |
7,050 | 11,502 | 4,452 | 63% | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total operating expenses |
26,317 | 61,889 | 35,572 | 135% | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Loss from operations |
(26,317 | ) | (61,889 | ) | (35,572 | ) | 135% | |||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Other income (expense), net |
||||||||||||||||
| Interest income |
1,524 | 1,362 | (162 | ) | (11% | ) | ||||||||||
| Interest expense – related party |
— | (1,433 | ) | (1,433 | ) | 100% | ||||||||||
| Other income (expense) |
7 | (15 | ) | (22 | ) | (314% | ) | |||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total other income (expense), net |
1,531 | (86 | ) | (1,617 | ) | (106% | ) | |||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Net loss |
$ | (24,786 | ) | $ | (61,975 | ) | $ | (37,189 | ) | 150% | ||||||
|
|
|
|
|
|
|
|
|
|||||||||
Operating Expenses
Research and Development Expenses
The following table summarizes our research and development expenses for the periods indicated (in thousands, except percentages):
| YEAR ENDED DECEMBER 31, |
CHANGE | |||||||||||||||
| 2024 | 2025 | $ | % | |||||||||||||
| Direct program costs: |
||||||||||||||||
| Ipsoprubart |
$ | 6,794 | $ | 26,100 | $ | 19,306 | 284% | |||||||||
| Other programs |
4,360 | 12,594 | 8,234 | 189% | ||||||||||||
| Unallocated costs: |
||||||||||||||||
| Personnel-related |
6,115 | 9,339 | 3,224 | 53% | ||||||||||||
| Facilities, overhead and other |
1,998 | 2,354 | 356 | 18% | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total research and development |
$ | 19,267 | $ | 50,387 | $ | 31,120 | 162% | |||||||||
|
|
|
|
|
|
|
|
|
|||||||||
Research and development expenses increased from $19.3 million for the year ended December 31, 2024 to $50.4 million for the year ended December 31, 2025. The $31.1 million increase was primarily attributable to the following:
| ∎ | a $19.3 million increase in expenses related to our ipsoprubart program, primarily driven by higher clinical research organization (CRO) costs, contract manufacturing organization (CMO) costs, personnel-related expenses and consulting fees associated with the advancement of ipsoprubart into its Phase 2/3 clinical trial, including site initiation and patient enrollment activities that began in 2025; |
| ∎ | an $8.2 million increase in expenses related to our other development programs, primarily due to higher CMO costs, material costs, and consulting fees incurred in programs unrelated to ipsoprubart; |
| ∎ | a $3.2 million increase in personnel-related expenses, primarily attributable to increased headcount to support the advancement of our research and development activities; and |
| ∎ | a $0.4 million increase in other research and development expenses, primarily driven by higher rent expense and costs associated with increased headcount. |
99
Table of Contents
General and Administrative Expenses
The following table summarizes our general and administrative expenses for the periods indicated (in thousands, except percentages):
| YEAR ENDED DECEMBER 31, |
CHANGE | |||||||||||||||
| 2024 | 2025 | $ | % | |||||||||||||
| Personnel-related |
$ | 2,834 | $ | 3,370 | $ | 536 | 19 | % | ||||||||
| Third-party service providers |
3,628 | 7,178 | 3,550 | 98 | % | |||||||||||
| Facilities, overhead and other |
588 | 954 | 366 | 62 | % | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total general and administrative |
$ | 7,050 | $ | 11,502 | $ | 4,452 | 63 | % | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
General and administrative expenses increased from $7.1 million for the year ended December 31, 2024 to $11.5 million for the year ended December 31, 2025. The $4.5 million increase was primarily attributable to the following:
| ∎ | a $3.6 million increase in fees for third-party service providers, primarily due to higher legal and accounting expenses incurred in relation to financing activities; |
| ∎ | a $0.5 million increase in personnel-related expenses, primarily driven by increased general and administrative headcount to support our growth; and |
| ∎ | a $0.4 million increase in other general and administrative expenses, primarily driven by higher rent expense and costs associated with increased headcount. |
Other Income (Expense), Net
Interest Income
Interest income decreased from $1.5 million for the year ended December 31, 2024 to $1.3 million for the year ended December 31, 2025. The $0.2 million, or 11%, decrease was primarily due to lower interest rates.
Interest Expense
Interest expense was $1.4 million for the year ended December 31, 2025 and related to the difference between the proceeds received from convertible promissory notes issued in March and June 2025 and the fair value of the notes, which was due to accrued interest on the notes as of the conversion date. There was no interest expense during the year ended December 31, 2024 as our convertible promissory notes were issued after that fiscal year.
Other Income (Expense)
Other expense, net was less than $0.1 million for each of the years ended December 31, 2024 and 2025 and related primarily to foreign currency gains and losses.
Liquidity and Capital Resources
Sources of Liquidity
Since our inception, we have incurred significant net losses and negative cash flows in each period and on an aggregate basis. We have incurred significant net operating losses from operations since inception and had an accumulated deficit of $173.0 million as of December 31, 2025. Additionally, we anticipate that our operating losses and negative operating cash flows will increase for the foreseeable future as we continue to expand our research and development programs.
Since our inception, we have devoted substantially all of our resources to organizing and staffing our company, business planning, identifying product candidates, establishing our intellectual property portfolio, conducting research and nonclinical studies, including IND-enabling studies, initiating and conducting clinical trials, establishing arrangements with third parties for the manufacture of our product candidates, and providing general and administrative support for these operations.
100
Table of Contents
To date, we have financed our operations primarily through the issuance and sale of shares of our redeemable convertible preferred stock and issuance of convertible promissory notes. As of December 31, 2025, issuances and sales of our redeemable convertible preferred stock have resulted in aggregate gross proceeds of $251.4 million.
On March 7, 2025, we issued convertible promissory notes with an aggregate principal amount of $20.0 million to multiple investors.
On June 30, 2025, we issued additional convertible promissory notes with an aggregate principal amount of $20.0 million.
On October 15, 2025, we issued 9,134,315 shares of our Series C redeemable convertible preferred stock at a purchase price of $10.15 per share, resulting in gross proceeds of $92.8 million. Concurrently, the outstanding principal of all our convertible promissory notes and all accrued and unpaid interest thereon, totaling $41.4 million, automatically converted into 4,080,265 shares of our Series C redeemable convertible preferred stock at a conversion price of $10.15 per share. The aggregate gross proceeds from the total of 13,214,580 shares either newly issued or converted from convertible promissory notes in this first closing was $132.8 million.
On June 18, 2026, we completed the second closing of our Series C redeemable convertible preferred stock financing and issued 4,918,477 shares of our Series C redeemable convertible preferred stock at a purchase price of $10.15 per share, resulting in gross proceeds of $49.9 million.
Funding Requirements
As of December 31, 2025, we had total cash, cash equivalents and marketable securities of $86.5 million. In addition, on June 18, 2026, we completed the second closing of our Series C redeemable convertible preferred stock financing and issued 4,918,477 shares of our Series C redeemable convertible preferred stock at a purchase price of $10.15 per share, resulting in gross proceeds of $49.9 million. Based on our current operating plan, we estimate that our existing cash, cash equivalents and marketable securities as of the date of this prospectus, together with the estimated net proceeds from this offering, will be sufficient to fund our projected operating expenses and capital expenditures into 2029. However, this estimate is a forward-looking statement that involves risks and uncertainties, and actual results could vary materially. We have based this estimate on assumptions that may prove to be wrong, and we could deplete our capital resources sooner than we expect.
We have incurred recurring losses and negative cash flows from operations and expect to continue to incur losses for the foreseeable future. Without giving effect to the anticipated net proceeds from this offering, we estimate that our existing cash, cash equivalents and marketable securities as of December 31, 2025 and the proceeds from the issuance of the second tranche of the Series C convertible preferred stock in June 2026 will not be sufficient to fund our operations for the next 12 months and, therefore, have concluded that circumstances exist that raise substantial doubt about our ability to continue as a going concern.
Our principal uses of cash in recent periods have been funding our research and development activities, including personnel-related costs, and general and administrative expenses. Our future capital requirements will depend on many factors, including, but not limited to:
| ∎ | the scope, progress, results and costs of researching, developing and manufacturing our product candidates or any future product candidates, and conducting nonclinical studies; |
| ∎ | the number and characteristics of any additional product candidates we develop or acquire; |
| ∎ | the timing of, and the costs involved in, obtaining regulatory approvals or clearances for our product candidates or any future product candidates; |
| ∎ | the expenses needed to attract and retain skilled personnel; |
| ∎ | the cost of any future product candidates and any products we successfully commercialize; |
| ∎ | the timing, receipt and amount of sales of any future approved or cleared products, if any; and |
| ∎ | our ability to establish and maintain strategic collaborations, licensing or other arrangements and the financial terms of any such agreements that we may enter into, including the timing and amount of any future milestone, royalty or other payments due under any such agreement. |
101
Table of Contents
We may not be able to raise additional funds or enter into such other arrangements when needed or on favorable terms, or at all. To the extent that we raise additional capital through the sale of equity or convertible debt securities, the ownership interest of our stockholders will be or could be diluted, and the terms of these securities may include liquidation or other preferences that adversely affect the rights of our common stockholders. Debt financing and equity financing, if available, may involve agreements that include covenants limiting or restricting our ability to take specific actions, such as incurring additional debt, making capital expenditures or declaring dividends. If we raise additional funds through future collaboration or licensing arrangements with third parties or other strategic transactions, we may have to relinquish rights to our intellectual property, future revenue streams, research programs or product candidates, or grant licenses on terms that may not be favorable to us and/or may reduce the value of our common stock. If we are unable to obtain adequate funding as and when needed, or on attractive terms, we could be required to significantly delay, reduce, or eliminate some or all of our research and development activities, product portfolio expansion or commercialization efforts, out-license intellectual property rights to our product candidates, sell unsecured assets, or scale back or terminate our pursuit of new strategic arrangements and transactions, or a combination of the above, any of which may have a material adverse effect on our business, results of operations, and financial condition.
Cash Flow Summary
The following table summarizes our cash flows for the periods presented (in thousands):
| YEAR ENDED DECEMBER 31, |
||||||||||||
| 2024 | 2025 | CHANGE | ||||||||||
| Net cash provided by (used in): |
||||||||||||
| Operating activities |
$ | (26,003 | ) | $ | (60,484 | ) | $ | (34,481 | ) | |||
| Investing activities |
29,701 | (52,978 | ) | (82,679 | ) | |||||||
| Financing activities |
— | 131,683 | 131,683 | |||||||||
|
|
|
|
|
|
|
|||||||
| Net increase in cash and cash equivalents |
$ | 3,698 | $ | 18,221 | $ | 14,523 | ||||||
|
|
|
|
|
|
|
|||||||
Operating Activities
Net cash used in operating activities for the year ended December 31, 2024 of $26.0 million was primarily due to our net loss of $24.8 million, $2.8 million net change in operating assets and liabilities, and $0.7 million of net amortization of premiums and discounts of marketable securities, partially offset by $2.3 million non-cash stock-based compensation expense.
Net cash used in operating activities for the year ended December 31, 2025 of $60.5 million was primarily due to our net loss of $62.0 million and $1.7 million of net change in operating assets and liabilities, adjusted for $1.8 million non-cash stock-based compensation expense and $1.4 million non-cash interest expense related to convertible promissory notes.
Investing Activities
Net cash provided by investing activities was $29.7 million for the year ended December 31, 2024, which consisted primarily of $34.5 million of proceeds from the maturity of marketable securities, partially offset by $4.8 million in purchases of marketable securities.
Net cash used in investing activities was $53.0 million for the year ended December 31, 2025, which consisted primarily of the purchase of marketable securities.
Financing Activities
Net cash provided by financing activities was $131.7 million for the year ended December 31, 2025, which consisted primarily of net proceeds from the issuance and sale of shares of our Series C redeemable convertible preferred stock and issuance of our convertible promissory notes, which were fully converted during the year ended December 31, 2025 into shares of our Series C redeemable convertible preferred stock as described in greater detail above.
102
Table of Contents
There were no financing activities during the year ended December 31, 2024.
Contractual Obligations and Other Commitments
We enter into contracts in the normal course of business with various third parties for nonclinical studies, clinical trials, manufacturing services and other services. These contracts generally provide for termination upon limited notice, and therefore we believe that our noncancelable obligations under these agreements are not material. All contracts are terminable, with varying provisions regarding termination. If a contract with a specific vendor were to be terminated, we would only be obligated for the products or services that we had received through the time of termination.
In September 2025, we entered into a noncancelable operating lease agreement for approximately 14,055 rentable square feet of office and laboratory space in South San Francisco, California. The lease has a three-year term expiring on September 30, 2028, and includes an initial rent abatement period. The lease provides us with two consecutive options to extend the lease term for three years each, at the then-prevailing fair rental value. We have not included these extension options in the lease term as it is not reasonably certain to exercise either option. The lease provides for escalating annualized base rent payments starting at $0.9 million and increasing to $1.0 million in the final year of the lease. Remaining lease payments from January 1, 2026 through the end of the lease term total $2.3 million. For additional details, see Note 7 to our financial statements included elsewhere in this prospectus.
We are also party to a mouse platform agreement with an unrelated third party. Under the agreement, we were granted non-exclusive licenses to use transgenic mice to generate antibodies and to exploit products containing the resulting antibodies (Antibody Products). Ipsoprubart (but not ELA822) was generated under this agreement. We own all intellectual property rights in the antibodies generated from our use of the mice and in all Antibody Products.
Under the agreement, we paid a one-time initial technology access fee and are obligated to pay annual low five-digit maintenance fees per distinct antigen with which we immunized the mice, continuing until the first commercial sale of an Antibody Product. In addition, the agreement requires us to make certain development and commercial milestone payments upon the achievement of specified events by each Antibody Product, as well as pay royalties on net sales of Antibody Products. Aggregate milestone payments are up to the low double-digit millions of dollars per distinct Antibody Product. Royalties under the agreement are a low single digit percentage of net sales of all Antibody Products and are payable on a product-by-product and country-by-country basis beginning with the first commercial sale of the applicable Antibody Product in the applicable country and continuing for 10 years thereafter. The agreement remains in effect until the expiration of the last-to-expire royalty term, unless earlier terminated by us at will or by either party for the other party’s uncured material breach of a material provision of the agreement.
Off-Balance Sheet Arrangements
Since our inception, we have not had, and we do not currently have, any off-balance sheet arrangements as defined in the rules and regulations of the SEC.
Critical Accounting Estimates
The preparation of these financial statements in accordance with accounting principles generally accepted in the United States (U.S. GAAP) requires us to make estimates and assumptions for the reported amounts of assets, liabilities, revenue, expenses and related disclosures. Our estimates are based on our historical experience and on various other factors that we believe are reasonable under the circumstances, the results of which form the basis for making judgments about the carrying value of assets and liabilities that are not readily apparent from other sources. Actual results may differ from these estimates under different assumptions or conditions and any such differences may be material.
We consider an accounting estimate to be critical if: (i) the accounting estimate requires us to make assumptions about matters that were highly uncertain at the time the accounting estimate was made, and (ii) changes in the estimate that are reasonably likely to occur from period to period, or use of different estimates that we reasonably could have used in the current period, would have a material impact on our financial condition or results of operations.
103
Table of Contents
Management concluded there were three critical accounting estimates as of and for the year ended December 31, 2025: accrued research and development expenses, stock-based compensation, and common stock valuations.
Accrued Research and Development Expenses
We estimate accrued research and development expenses based on the services performed, pursuant to contracts with research institutions and third-party service providers that conduct and manage nonclinical studies, clinical trial manufacturing services and other research services on our behalf. We record the costs of research and development activities based on the estimated services provided but not yet invoiced and include these costs in accrued expenses in the balance sheet. These costs are a component of our research and development expenses.
We accrue these costs based on factors such as estimates of the work completed in accordance with agreements established with our third-party service providers. We make judgments and estimates in determining the accrued expenses balance. As actual costs become known, we adjust our accrued expenses. We have not experienced any material differences between accrued costs and actual costs incurred. However, the status and timing of actual services performed may vary from our estimates, leading to adjustments to expenses in future periods. Changes in these estimates that result in material changes to our accrued expenses could materially affect our results of operations.
Stock-Based Compensation
We estimate the fair value of stock options on the grant date using the Black-Scholes option pricing model, which requires us to make a number of assumptions considering, among other things, contemporaneous valuations of our common stock prepared by independent third-party valuation firms. The grant date fair value of stock-based awards is generally recognized on a straight-line basis over the requisite service period, which is generally the vesting period of the awards.
The Black-Scholes option pricing model requires the use of subjective assumptions to determine the fair
value of stock-based awards, including:
| ∎ | Fair Value of Common Stock–See the subsection titled “—Common Stock Valuations” below. |
| ∎ | Expected Term–We determined the expected term of our stock options using the simplified method. The simplified method calculates the expected term as the average of the time-to-vesting and the contractual life of the stock options. |
| ∎ | Expected Volatility–We estimated expected volatility data based on a study of publicly traded industry peer companies. To identify these peer companies, we considered the industry, stage of development, size, and financial leverage of potential comparable companies. Historical volatility is calculated based on a period of time commensurate with the expected term assumption. |
| ∎ | Expected Dividend–Our assumption for the expected dividend yield is zero as we have no history or expectation of paying cash dividends on our common stock in the foreseeable future. |
| ∎ | Risk-Free Interest Rate–We based the risk-free interest rate on the yield available on U.S. Treasury zero-coupon issues similar in duration to the expected term of the award. |
See Note 11 to our financial statements included elsewhere in this prospectus for information concerning certain of the specific assumptions we used in applying the Black-Scholes option pricing model to determine the estimated fair value of our stock options granted in the period presented. Such assumptions involve inherent uncertainties and the application of significant judgment. As a result, if factors or expected outcomes change and we use significantly different assumptions or estimates, our stock-based compensation expense could be materially different.
The aggregate intrinsic value of all outstanding stock options as of June 30, 2026, was $57.5 million based on the assumed initial public offering price of $15.00 per share, which is the midpoint of the price range set forth on the cover of this prospectus, of which approximately $16.0 million was related to vested stock options and approximately $41.5 million was related to unvested stock options.
Common Stock Valuations
Our stock-based compensation includes stock options issued under the 2022 Stock Plan and profits interest units which were granted under Electra Therapeutics LLC’s 2023 Equity Incentive Plan. In the six months ended June 30,
104
Table of Contents
2026, we recorded stock-based compensation expenses of $2.4 million related to stock options and $0.2 million related to profits interest units. No profits interest units have been granted by Electra Therapeutics LLC subsequent to February 2024.
For the grant of stock options, as there has been no public market of our common stock and preferred stock to date, the estimated fair value of our common stock has been determined by our board of directors with input from management and with consideration of independent third party valuations. Given the absence of a public market for our common stock, our board of directors exercised reasonable judgment and considered a number of objective and subjective factors to determine the best estimate of the fair market value, including: our stage of development and material risks related to our business; the progress of our research and development programs; the prices at which we sold shares of our redeemable convertible preferred stock; the rights, preferences and privileges of our redeemable convertible preferred stock relative to those of common stock; our financial condition and operating results, including our levels of available capital resources; external market conditions in the life sciences industry; general U.S. market and economic conditions; equity market conditions of comparable public companies; the lack of marketability of our securities; and the likelihood of achieving a liquidity event such as an initial public offering (IPO) in light of prevailing market conditions. Valuations of our common stock were prepared in accordance with the guidance outlined in the American Institute of Certified Public Accountants’ Accounting and Valuation Practice Guide, Valuation of Privately-Held-Company Equity Securities Issued as Compensation.
Consistent with the guidance, for the valuations performed to date, we determined the hybrid method was the most appropriate method for allocating the equity value to the outstanding classes of securities. The hybrid method uses a hybrid approach between the option pricing method (OPM) and a probability-weighted expected return method (PWERM). The PWERM is a scenario-based analysis that estimates value per share based on the probability-weighted present value of expected future investment returns, considering each of the possible outcomes available to us, as well as the economic and control rights of each share class. Under the hybrid method, the equity value is estimated based on probability weighted value across multiple scenarios where the OPM is used to estimate the allocation of value within one or more of those scenarios. In our application of the hybrid method, we assumed a liquidity scenario of an IPO and a stay-private scenario, and our equity value was probability weighted across those scenarios. Our board of directors also considered the fact that our common stock could not freely trade in the public markets and applied appropriate discounts for lack of marketability.
Once a public trading market for our common stock has been established in connection with the completion of this offering, it will no longer be necessary for our board of directors to estimate the fair value of our common stock in connection with our accounting for granted stock options and other such awards we may grant, as the fair value of our common stock will be determined based on the quoted market price of our common stock. Further expense amounts for any particular period could be affected by changes in our assumptions or market conditions.
Recent Accounting Pronouncements
See Note 2 to our financial statements included elsewhere in this prospectus.
Emerging Growth Company and Smaller Reporting Company Status
We expect to be an emerging growth company, as defined in the JOBS Act. Under the JOBS Act, emerging growth companies can delay adopting new or revised accounting standards issued subsequent to the enactment of the JOBS Act until such time as those standards apply to private companies. We have elected to use this extended transition period for complying with new or revised accounting standards that have different effective dates for public and private companies until the earlier of the date we (i) are no longer an emerging growth company or (ii) affirmatively and irrevocably opt out of the extended transition period provided in the JOBS Act. As a result, our financial statements may not be comparable to companies that comply with new or revised accounting pronouncements as of public company effective dates.
We are also a “smaller reporting company” as defined in the Exchange Act. We may continue to be a smaller reporting company even after we are no longer an emerging growth company. We may take advantage of certain of the scaled disclosures available to smaller reporting companies and will be able to take advantage of these scaled disclosures for so long as the market value of our voting and non-voting common stock held by non-affiliates is less
105
Table of Contents
than $250.0 million measured on the last business day of our second fiscal quarter, or our annual revenue is less than $100.0 million during the most recently completed fiscal year and the market value of our voting and non-voting common stock held by non-affiliates is less than $700.0 million measured on the last business day of our second fiscal quarter.
Quantitative and Qualitative Disclosures About Market Risk
Interest Rate and Market Risk
We are exposed to market risks in the ordinary course of our business, primarily limited to interest rate fluctuations. Financial instruments that potentially subject us to concentrations of credit risk primarily consist of cash, cash equivalents and marketable securities. We may maintain deposits in federally insured financial institutions in excess of federally insured limits and hold investments in money market funds and U.S. Treasury and corporate securities, which can be subject to certain credit risks. The primary objective of our investment activities is to preserve principal and liquidity while maximizing income without significantly increasing risk. We do not enter into investments for trading or speculative purposes.
Due to the short-term nature of our investment portfolio, we do not believe that a hypothetical 10% increase or decrease in interest rates during any of the periods presented would have a material effect on our financial statements included elsewhere in this prospectus.
Effects of Inflation
Inflation generally affects us by increasing our cost of labor and research and development contract costs. We do not believe inflation has had a material effect on our results of operations during the periods presented in our financial statements included elsewhere in this prospectus.
106
Table of Contents
Overview
We are a late clinical-stage biopharmaceutical company focused on pioneering a new class of precision medicines for the treatment of immune-mediated diseases and cancer. Our novel approach targets signal regulatory proteins (SIRP), a family of cell surface receptors whose expression is restricted to specific immune cell populations and increases upon activation, to enable selective depletion of disease-driving cells while preserving normal immune function. By replacing broad immunosuppression with selective elimination of principal cells that drive disease, we believe our approach can do for immune-mediated diseases what precision oncology has done for cancer, transforming the treatment paradigm for patients. To our knowledge, we are the first company to advance this SIRP-targeted precision immune cell depletion approach into clinical development and demonstrate proof-of-concept in humans.
Our lead product candidate is ipsoprubart, a novel pan-SIRP monoclonal antibody designed to selectively deplete pathological myeloid cells and T cells via binding to SIRPa/ß1/g, which is currently in a global registrational program for patients with secondary hemophagocytic lymphohistiocytosis (sHLH). In our Phase 1b trial, ipsoprubart was generally well tolerated and demonstrated a 100% 8-week overall survival (OS) rate and 100% overall response rate (ORR) in 12 frontline patients with malignancy-associated HLH (mHLH), the largest subset of sHLH and the population associated with the poorest outcomes. Of the 12 mHLH frontline patients, 10 achieved a partial response (PR) and two achieved a modified complete response (mCR). We are conducting SURPASS, our global Phase 2/3 registrational trial in newly diagnosed, treatment-naïve sHLH patients, as well as COMPASS, our natural history study designed to provide an external control comparator for SURPASS. We expect to complete enrollment in our registrational program in the second half of 2027. Ipsoprubart has received Breakthrough Therapy designation (BTD) from the U.S. Food and Drug Administration (FDA) and PRIority MEdicine (PRIME) designation from the European Medicines Agency (EMA), each granted for the treatment of sHLH broadly.
In addition, in eight evaluable sHLH patients with underlying T or B cell lymphomas treated across our Phase 1b trial and Expanded Access Program, who had tumor response measurements within weeks of treatment, ipsoprubart demonstrated a 100% objective tumor response rate, with seven out of eight patients achieving a complete response (CR). We are conducting a Phase 1 clinical trial evaluating ipsoprubart as a monotherapy in patients with relapsed/refractory T cell and natural killer (NK) cell malignancies, with initial data expected in the second half of 2027.
We are also advancing ELA822, a novel SIRPg-specific monoclonal antibody designed to selectively deplete activated T cells, with potential applications across chronic T cell-mediated immune and inflammatory diseases. In August 2026, we obtained regulatory clearance and initiated a Phase 1 clinical trial of ELA822 in healthy volunteers in Europe, with data in expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027, subject to regulatory approval.
107
Table of Contents
We have generated a proprietary library of SIRP-targeted antibodies with distinct profiles, developed over years of dedicated discovery and engineering. This platform provides substantial flexibility to align antibody design with the cellular biology and therapeutic objectives of each program, and we believe it positions us to pursue a broad range of indications involving pathogenic SIRP-expressing cells without requiring additional de novo discovery efforts. From our proprietary library of SIRP-targeted antibodies, we have advanced two product candidates, ipsoprubart and ELA822, each designed for distinct disease settings, as shown in the chart below:
Ipsoprubart for the Treatment of Secondary Hemophagocytic Lymphohistiocytosis (sHLH)
Ipsoprubart (formerly known as ELA026), our lead product candidate, is a novel pan-SIRP antibody engineered with an enhanced-effector Fc domain for rapid, potent depletion of pathogenic myeloid cells and T cells. We are developing ipsoprubart initially for sHLH, a severe hyperinflammatory syndrome with no broadly approved therapies and consistently poor survival over the past two decades. In mHLH, which accounts for approximately 45% of all sHLH cases and is associated with the poorest outcomes, historical 2-month OS rates with existing treatments are approximately 50%. Patients with sHLH are typically diagnosed and treated by hematologist-oncologists in both outpatient and inpatient settings. The diagnosed incidence in the United States reached approximately 12,600 in 2025, reflecting an 11.5% compound annual growth rate since 2021, and would reach nearly 17,500 in 2028 assuming a similar annual growth rate. We believe this estimate is conservative as it excludes misdiagnosed, undiagnosed, and non-International Classification of Diseases (ICD) (standardized diagnosis codes used to classify medical conditions) coded patients. As disease awareness increases, underlying triggering conditions become more prevalent, and diagnostic approaches become simpler, we believe the diagnosed incidence will continue to rise.
In our completed Phase 1b trial, ipsoprubart was generally well tolerated and achieved 100% OS at 8 weeks in 12 frontline mHLH patients, alongside a 100% ORR, rapid improvement of key pharmacodynamic (PD) biomarkers, and a 100% hospital discharge rate in all 11 patients evaluable for discharge (one patient withdrew prior to discharge assessment). Additional clinical experience beyond mHLH supports the potential of ipsoprubart as a treatment for sHLH broadly. Based on these data, ipsoprubart has received BTD from the FDA and PRIME designation from the EMA, each granted for the treatment of sHLH broadly. We initiated a global registrational program for ipsoprubart in sHLH consisting of SURPASS, a Phase 2/3 registrational trial, and COMPASS, our natural history study designed to provide an external control comparator for SURPASS, with enrollment completion expected in the second half of 2027.
Ipsoprubart for the Treatment of T/NK Cell Malignancies
In eight patients treated across our Phase 1b trial and Expanded Access Program, who had tumor response measurements within weeks of treatment, ipsoprubart achieved a 100% objective tumor response rate, including an 88% CR rate. One patient with heavily refractory T cell lymphoma treated with ipsoprubart as a monotherapy experienced a CR with respect to both sHLH and the underlying T cell lymphoma. The patient was alive as of the last follow-up, more than one year post treatment. T/NK cell malignancies remain areas of significant unmet medical need, particularly in the relapsed/refractory setting where available therapies demonstrate low CR rates, limited durability, and poor long-term outcomes. We believe these diseases represent a substantial commercial opportunity,
108
Table of Contents
and we estimate a diagnosed incidence rate of more than 13,000 diagnosed incidence annually in the United States across the T/NK cell malignancy subtypes. Based on the anti-tumor activity observed in the patients from the Phase 1b trial and Expanded Access Program, we have initiated a Phase 1 clinical trial of ipsoprubart in relapsed/refractory T/NK cell malignancies and expect initial data in the second half of 2027.
ELA822 for the Treatment of Chronic Immune and Inflammatory Diseases
ELA822, our second product candidate, is a novel SIRPg-specific antibody designed to selectively deplete activated, pathogenic T cells while sparing naïve and regulatory myeloid cells and T cells – a target profile we believe is well suited for a broad range of chronic T cell-mediated immune and inflammatory diseases where durable efficacy and long-term tolerability are important therapeutic objectives. Nonclinical data for ELA822 demonstrated selective depletion of activated T cells and in vivo activity in humanized models of T cell-driven diseases, including giant cell arteritis (GCA) and graft-versus-host disease (GvHD). ELA822 was well tolerated in non-human primate (NHP) studies. We have obtained regulatory clearance and initiated a Phase 1 clinical trial of ELA822 in healthy volunteers in Europe, with data expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027, subject to regulatory approval.
Our Approach: Precision SIRP-Targeted Immune Cell Depletion
The Unmet Need in Immune-Mediated Diseases
Pathogenic myeloid cells and T cells are key drivers of immune dysregulation across a broad range of diseases, including acute hyperinflammatory syndromes, chronic inflammatory conditions, and malignancies. Aberrant activation of these cells can lead to excessive release of numerous proinflammatory cytokines and mediators, driving tissue damage, systemic inflammation, and end-organ dysfunction. Current therapies are often limited either by broad immunosuppression associated with non-selective immune modulation or by insufficient disease control resulting from narrowly targeting individual cytokines.
We believe that selectively eliminating activated pathogenic cells, while preserving host defense and immune homeostasis, may provide a more effective therapeutic approach for diseases driven by dysregulated myeloid cell and T cell activity.
SIRP Expression as a Marker of Immune Cell Activation
Our approach leverages the unique biology of SIRP to enable selective depletion of activated pathogenic cells. SIRP is a family of cell surface receptors involved in regulating immune homeostasis and inflammatory responses. Their expression is largely restricted to defined immune cell populations and can increase significantly as these cells become activated. This activation-dependent expression pattern creates a molecular marker, or flag, that can be targeted to selectively deplete disease-driving cells while sparing naïve and regulatory immune populations necessary for normal immune function.
The principal members of the SIRP family include SIRPa and SIRPß1, expressed on dendritic cells and antigen-presenting cells (APCs), monocytes, macrophages, and granulocytes; and SIRPg, expressed predominantly on T cells, as shown in the figure below:
109
Table of Contents
Our Distinct Approach to Precision SIRP-Targeted Cell Depletion
Historically, drug development involving SIRP has focused primarily on disrupting the CD47-SIRPa interaction, an innate immune checkpoint commonly referred to as a “don’t eat me” signaling pathway that may be exploited by cancer cells to evade immune surveillance. These approaches have largely been pursued in oncology indications and, to date, have demonstrated limited monotherapy activity and encountered clinical development challenges.
Our approach differs fundamentally from these prior efforts. Rather than disrupting the CD47-SIRPa signaling, we seek to preserve this pathway to maintain immune surveillance while leveraging the lineage-restricted and activation-dependent expression pattern of SIRP to selectively deplete activated pathogenic cells.
Our antibodies are designed to selectively bind SIRP-expressing cells without blocking or otherwise modulating CD47-SIRPa signaling. Our approach is to mediate therapeutic activity through antibody-mediated elimination of the bound cell, achieved through antibody-dependent cellular cytotoxicity (ADCC) and antibody-dependent cellular phagocytosis (ADCP), resulting in elimination of the targeted pathogenic cell. Selectivity is enabled by differential SIRP expression between activated cells and naïve or regulatory immune cell populations. To our knowledge, we are the first company to advance this SIRP-targeted precision immune cell depletion approach into clinical development and demonstrate proof-of-concept in humans.
Our Strengths and Capabilities
We have built differentiated capabilities spanning SIRP-targeted antibody discovery and the underlying SIRP biology, resulting in our current pipeline. Together, we believe the following factors will enable us to achieve our goal of developing, commercializing, and maximizing the impact of precision immune cell depletion therapies across multiple disease areas:
Proprietary and Expansive SIRP-Targeted Antibody Library with Broad Therapeutic Optionality
We have generated a proprietary library of antibodies targeting members of the SIRP family through years of dedicated discovery efforts. This novel portfolio includes antibodies spanning a broad range of therapeutically relevant binding profiles, including pan-SIRP binders, selective binders targeting individual SIRP family members, and antibodies with differentiated epitope specificity, affinity, and effector-function characteristics.
Because SIRPa, SIRPß1, and SIRPg share substantial structural homology, selective targeting of individual SIRP family members has historically been challenging. Our discovery platform has produced antibodies with defined and differentiated binding characteristics, ranging from pan-SIRP engagement to highly specific SIRPa/SIRPß1 or SIRPg targeting.
We believe this library provides substantial flexibility to align antibody design with the cellular biology and therapeutic objectives of each program while creating barriers to entry based on proprietary know-how and intellectual property. We also believe it positions us to expand into additional indications involving pathogenic SIRP-expressing cells without requiring additional de novo antibody discovery efforts.
Deep Knowledge and Expertise in SIRP Biology
We have developed significant expertise in the biology of SIRP expression across activated immune cell populations and its potential application in precision immune cell depletion. Our work has characterized differential SIRP expression across myeloid cell and T cell subsets, which we believe can enable selective targeting of pathogenic cells while preserving immune populations important for normal immune function.
We have evaluated SIRP expression patterns relative to other immune cell markers used in targeted depletion approaches and believe SIRP expression may provide distinct therapeutic advantages in certain disease settings due to its activation-dependent characteristics.
In addition, because SIRPg is expressed only in primates and lacks a murine ortholog, preclinical characterization of SIRPg-targeted therapies has historically been limited. To address this limitation, we have developed humanized in vitro and in vivo systems designed to evaluate SIRPg engagement, depletion kinetics, and downstream effects on T cell biology in translationally relevant settings.
We are applying these capabilities across both our product candidates and earlier-stage discovery programs and believe they provide a differentiated foundation for the continued development of precision SIRP-targeted therapies.
110
Table of Contents
Proven Track Record of Drug Development and Commercialization in Immunology, Hematology and Oncology
We have assembled a seasoned leadership team with deep experience advancing novel mechanisms of action and first-in-class medicines from discovery through clinical development and commercial launch across the immunology, hematology, and oncology therapeutic settings that define our pipeline. Collectively, members of our team have contributed to the approval and commercialization of more than five first-in-class therapies that established new treatment paradigms for diseases with significant unmet need, as well as therapies that became among some of the most commercially successful product launches in the history of the pharmaceutical industry. Together, our experience spans translational science, clinical development, technical operations, biologics manufacturing, and commercial operations, which we believe provides us with the capabilities to advance innovative scientific concepts into therapies with meaningful patient impact and commercial potential.
Disciplined Research and Development Strategy and Efficient Execution for Advancing Novel Therapies
Our development strategy focuses on identifying targets and mechanisms with the potential to address underlying pathology across multiple diseases. Rather than pursuing targets applicable primarily to a single indication, we seek to connect target biology to disease pathology in order to prioritize indications with strong mechanistic rationale. Both of our product candidates are designed to address multiple immunology and oncology indications, with applicability determined by the specific immune cell population driving disease pathophysiology.
In addition, we prioritize opportunities with significant unmet need where meaningful clinical impact could translate into substantial commercial opportunity, such as our lead program in sHLH, where there are no broadly approved therapies and outcomes remain poor.
Our approach emphasizes rigorous translational evaluation and focused clinical execution in order to achieve clinical validation early in the program. In certain cases, this includes pursuing indications with limited development precedent or no approved therapies, which may require novel clinical and regulatory strategies, such as the use of external control comparator data from our COMPASS natural history study to support the registrational program.
Since our founding in 2018, we have used this development strategy to progress ipsoprubart from an initial scientific concept to a novel product candidate to the commencement of a global registrational program and expansion into additional indications. We next advanced ELA822, another novel product candidate for immune-mediated diseases, to CTA approval and have initiated a Phase 1 clinical trial of ELA822 in healthy volunteers in Europe, with data expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027, subject to regulatory approval.
Our Strategy
Our goal is to become a leading company in precision immune cell depletion with first-in-class or best-in-class SIRP-targeted therapies designed to transform treatment for patients with immune-mediated diseases and cancer. To achieve this goal, we intend to pursue the following:
| ∎ | Advance ipsoprubart through our registrational program as a novel treatment for patients with sHLH. sHLH is a rare, rapidly progressive hyperinflammatory disease with diagnosed incidence of approximately 12,600 in the United States in 2025, growing at an estimated 11.5% compound annual growth rate from 2021 through 2025. There are no approved therapies for sHLH broadly, and historical 2-month OS rates with existing treatments are approximately 50%. In our Phase 1b trial, ipsoprubart was generally well tolerated and demonstrated a 100% 8-week OS rate in 12 frontline patients with mHLH, as well as a 100% ORR and a 100% hospital-discharge rate in all 11 patients evaluable for discharge (one patient withdrew prior to discharge assessment). Ipsoprubart has received BTD from the FDA and PRIME designation from the EMA, in each case for sHLH broadly. We are conducting SURPASS, our global Phase 2/3 registrational trial in newly diagnosed, treatment-naïve sHLH patients, as well as COMPASS, our natural history study designed to provide an external control comparator for SURPASS. We expect to complete enrollment in the second half of 2027. Subject to positive results, we plan to pursue regulatory approval that may position ipsoprubart as the first therapy approved for sHLH broadly. |
| ∎ | Expand the therapeutic application of ipsoprubart across hematologic malignancies and other inflammatory indications. We believe the mechanism underlying ipsoprubart’s activity seen in sHLH, targeting activated |
111
Table of Contents
| myeloid cells and T cells, may have broad applicability across T and B cell malignancies, severe cytokine release syndrome associated with immunotherapies, and other hyperinflammatory syndromes. Across our Phase 1b trial and Expanded Access Program, eight sHLH patients with underlying T or B cell lymphomas achieved both an overall response in sHLH and a 100% objective tumor response. Seven of the eight patients achieved a CR within weeks of ipsoprubart treatment, with or without concurrent standard of care (SOC) therapy for the underlying malignancy. We are conducting a Phase 1 clinical trial evaluating ipsoprubart as monotherapy in patients with relapsed/refractory T and NK cell malignancies, with initial data expected in the second half of 2027. |
| ∎ | Build a focused and capital-efficient commercial organization, initially centered on ipsoprubart. Patients with sHLH are primarily treated by hematologist-oncologists, with approximately 64% of patients in the United States managed at academic medical centers. We believe this highly concentrated prescriber base supports an efficient commercial model for ipsoprubart. If approved, we intend to commercialize ipsoprubart independently in the United States through a focused commercial organization. |
| ∎ | Advance ELA822 and demonstrate proof-of-concept in clinical trials for chronic T cell-mediated immune and inflammatory diseases. ELA822 is designed to selectively bind SIRPg and deplete activated pathogenic T cells, while sparing naïve and regulatory T cells as well as myeloid cells. We believe this target profile may be well suited for chronic diseases in which long-term tolerability and efficacy are important therapeutic considerations. Nonclinical data generated to date for ELA822 have demonstrated selective depletion of activated T cells, in vivo activity in humanized models of T cell-driven disease, subcutaneous bioavailability supportive of convenient dosing, and was well tolerated in NHP studies. In August 2026, we obtained regulatory clearance and initiated a Phase 1 clinical trial of ELA822 in healthy volunteers in Europe, with data expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027, subject to regulatory approval. |
| ∎ | Expand our pipeline in SIRP-targeted precision immunology and oncology. Supported by our proprietary library of SIRP-targeted antibodies and expertise in SIRP biology, we intend to broaden the therapeutic application of our product candidates and expand our pipeline with additional programs. |
| ∎ | Selectively explore value-creating strategic collaborations to maximize the value of our programs. We intend to selectively explore strategic collaborations where we believe partner capabilities can enhance the development, manufacturing, and commercialization potential of our product candidates. Accordingly, we may enter into collaborations that provide access to complementary expertise, expand the reach of our approach, accelerate development timelines, and strengthen our capital resources, while allowing us to retain meaningful participation in the long-term value creation of our programs. |
Our Team and Investors
Our seasoned leadership team brings complementary expertise across all stages of drug development, spanning immunology, hematology, and oncology therapeutic areas. Collectively, members of our leadership team have contributed to the development and commercialization of more than five first-in-class therapies with track records at organizations including Star Therapeutics, True North Therapeutics, Sanofi, Gilead Sciences, Novartis, and Genentech. Our combined experience exemplifies our demonstrated ability to translate novel biological insights into therapeutic candidates and advance them from discovery to commercialization.
| ∎ | Kathy Dong, Pharm.D., M.B.A., President and Chief Executive Officer. Dr. Dong has more than two decades of experience spanning corporate development, operational leadership, portfolio management, and commercialization. Prior to joining Electra, she supported the advancement of programs from discovery through clinical development in senior leadership roles at biotechnology companies focused on immunology and hematology, including Star Therapeutics and True North Therapeutics. Earlier in her career, she led the launch of multiple therapies, including SOVALDI and HARVONI at Gilead Sciences. |
| ∎ | Kim-Hien Dao, D.O., Ph.D., Chief Medical Officer. Dr. Dao is a board-certified hematologist and clinician-scientist with broad experience spanning translational research, clinical medicine, and drug development in malignancies and hyperinflammatory diseases. Prior to joining Electra, she held leadership roles across academic medicine and biopharmaceutical organizations and most recently led clinical development of multiple programs at Astex Pharmaceuticals. |
112
Table of Contents
| ∎ | Graham Parry, Ph.D., Chief Scientific Officer. Dr. Parry brings extensive experience in research and development across the biopharmaceutical industry and academia. His prior scientific leadership roles have included programs focused on inflammatory, hematologic, and neurodegenerative diseases and most recently served as Executive Vice President of Translational Sciences at Star Therapeutics. |
| ∎ | Gary Koe, Ph.D., Chief Technical Officer. Dr. Koe brings more than 30 years of experience and deep expertise in biologics chemistry, manufacturing and controls (CMC). In his prior roles, he led the scale-up, manufacturing, and quality control development of multiple biologic therapies and most recently served as Head of Technical Operations at Star Therapeutics and Aravive. |
| ∎ | Chris Clark, C.F.A., Chief Financial Officer. Mr. Clark brings more than two decades of expertise in biopharmaceutical capital markets and corporate strategy. Prior to joining Electra, he was a Portfolio Manager at RS Investments, where he led biopharma investments across multiple fund strategies. Earlier in his career, he held roles at TIAA-CREF and Dresdner RCM. |
Since our inception, we have raised more than $300 million in equity capital from a syndicate of leading healthcare and life sciences investors. Potential investors should not consider investments made by our existing investors as a factor when making a decision to purchase shares in this offering, as our existing investors likely have different risk tolerances and paid significantly less per share than the price at which shares are being offered in this offering.
Ipsoprubart: Lead Product Candidate for the Treatment of sHLH
Ipsoprubart, our lead product candidate, is a novel pan-SIRP antibody engineered with an enhanced-effector Fc domain for rapid, potent depletion of pathogenic myeloid cells and T cells. We are developing ipsoprubart initially for sHLH, a severe hyperinflammatory syndrome with no broadly approved therapies and consistently poor survival over the past two decades. In mHLH, which accounts for approximately 45% of all sHLH cases and is associated with the poorest outcomes, historical 2-month OS rates with existing treatments are approximately 50%. Patients with sHLH are typically diagnosed and treated by hematologist-oncologists in both outpatient and inpatient settings. The diagnosed incidence in the United States reached approximately 12,600 in 2025, reflecting an 11.5% compound annual growth rate since 2021, and would reach nearly 17,500 in 2028 assuming a similar annual growth rate. We believe this estimate is conservative as it excludes misdiagnosed, undiagnosed, and non-ICD coded patients. As disease awareness increases, underlying triggering conditions become more prevalent, and diagnostic approaches become simpler, we believe diagnosed incidence will continue to rise.
In our completed Phase 1b trial, ipsoprubart was generally well tolerated and achieved 100% OS at 8 weeks in 12 frontline mHLH patients, alongside a 100% ORR, rapid improvement of key PD biomarkers, and a 100% hospital discharge rate in all 11 patients evaluable for discharge (one patient withdrew prior to discharge assessment). Additional clinical experience beyond mHLH supports the potential of ipsoprubart as a treatment for sHLH broadly. Based on these data, ipsoprubart received BTD from the FDA and PRIME designation from the EMA, each granted for the treatment of sHLH broadly. We initiated a global registrational program for ipsoprubart in sHLH consisting of SURPASS, a Phase 2/3 registrational trial, and COMPASS, our natural history study designed to provide an external control comparator for SURPASS, with enrollment completion expected in the second half of 2027.
Overview of sHLH: A Rapidly Progressive Hyperinflammatory Syndrome with High Mortality and No Broadly Approved Therapies
HLH is a rare, life-threatening hyperinflammatory disease characterized by immune dysregulation leading to excessive cytokine release and rapid progression to multiorgan dysfunction. HLH is broadly categorized into two forms: primary HLH (pHLH), which is caused by inherited genetic mutations and typically diagnosed in infants and young children, and sHLH, which may be triggered by malignancy, infection, autoimmune disease, or immune-activating therapies that precipitate immune dysregulation in susceptible individuals. We estimate sHLH represents over 95% of HLH cases in the United States.
The pathophysiology of sHLH is characterized by failure to terminate activation and proliferation of T cells and macrophages following a strong antigenic stimulus. Rather than resolving, this immune activation spirals. Activated immune cells release inflammatory cytokines, creating a self-amplifying inflammatory cycle that can overwhelm the body’s ability to maintain immune homeostasis. Hemophagocytosis, or the engulfment of healthy red blood cells, white blood cells, and platelets by activated macrophages, is often a defining feature of the disease and is now understood to be one consequence of the broader immune dysregulation.
113
Table of Contents
sHLH is considered a medical emergency as uncontrolled hyperinflammation can progress to multiorgan failure and death. sHLH may be triggered by a range of underlying conditions, generally grouped into the following subtypes:
| ∎ | mHLH, which accounts for approximately 45% of all sHLH cases. Lymphoma is the predominant malignancy trigger and is associated with the poorest outcomes among sHLH subtypes. |
| ∎ | Infection-associated HLH (iHLH), which is triggered by various viral and intracellular pathogens, such as EBV, CMV, HIV, and COVID-19. |
| ∎ | Autoimmune-associated HLH (aHLH) / Macrophage Activation Syndrome (MAS), which may occur in autoimmune diseases, such as systemic juvenile idiopathic arthritis and adult-onset Still’s disease. |
| ∎ | Immune Effector Cell-associated HLH-like Syndrome (IEC-HS), which is increasingly recognized in association with immune-effector cell therapies, including chimeric antigen receptor T cell (CAR-T) therapies and bispecific T cell engagers (TCEs), both of which are increasing in clinical use, including in diseases beyond cancer. |
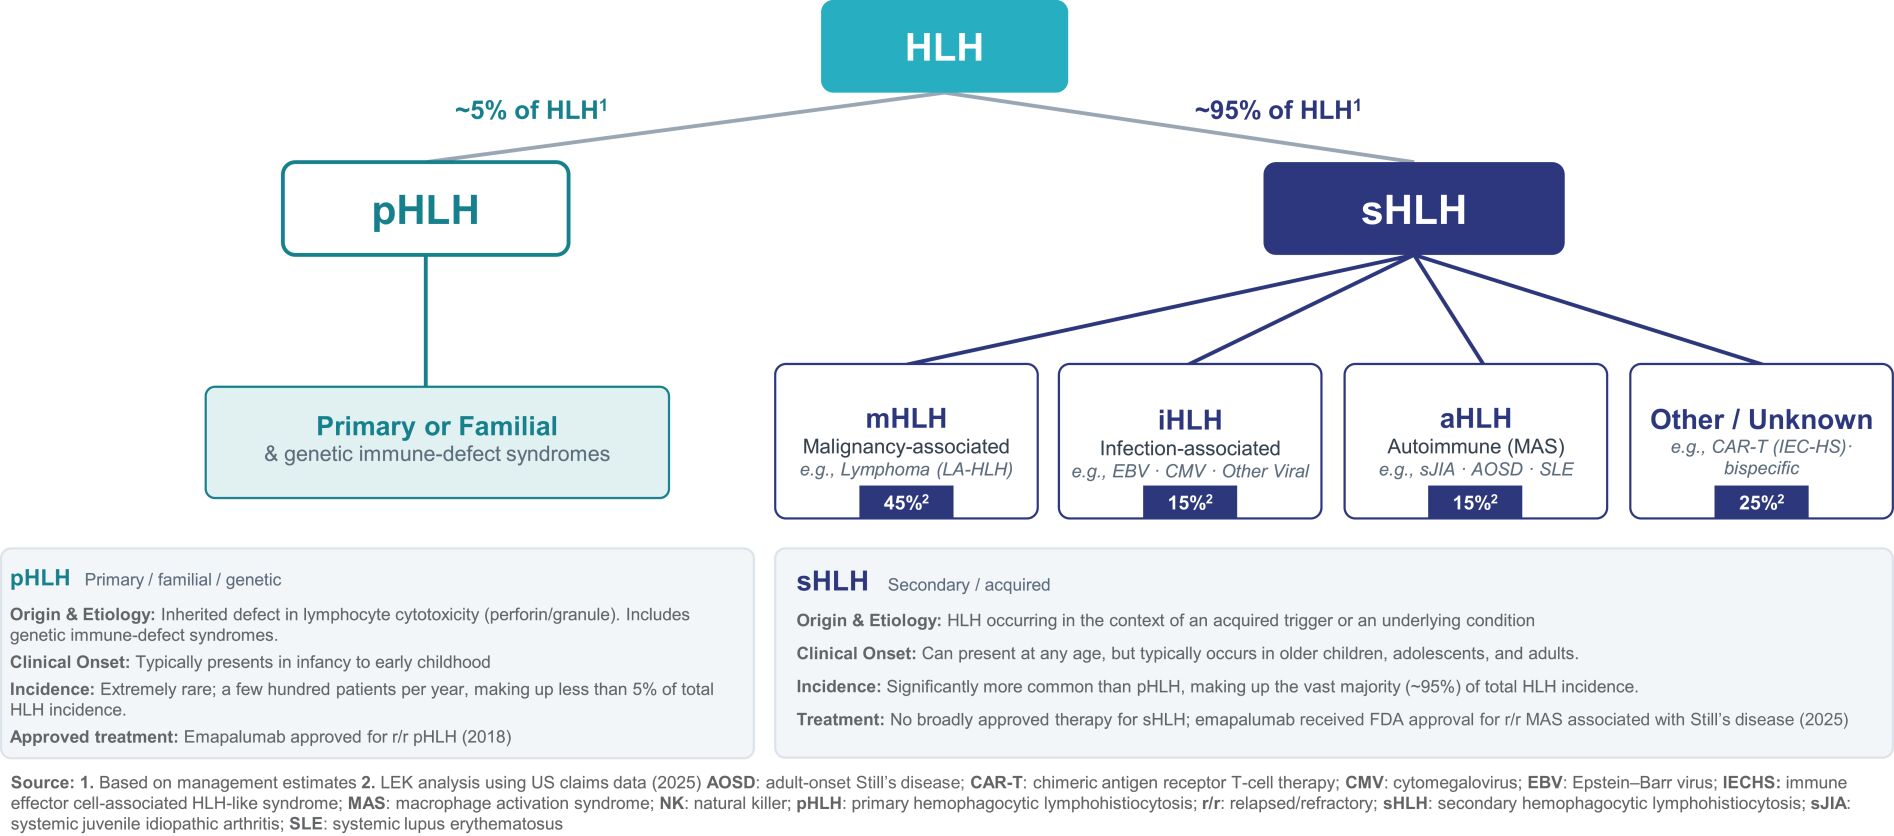
Across these etiologies, the underlying pathology is consistently that of dysregulated immune activation and proliferation of pathogenic T cells and macrophages resulting in systemic hyperinflammation.
Clinical Presentation and Diagnosis: Severe Hyperinflammation with Rapid Onset
The hyperinflammatory response associated with HLH leads to a rapid onset of severe clinical features. Over days to weeks, patients can develop a constellation of signs and symptoms that include persistent high fevers despite use of antipyretics or antibiotics, severe fatigue, abdominal pain and fullness from an enlarged spleen and liver, easy bruising or bleeding, and in some cases rash, jaundice, or neurologic changes such as confusion. Patients feel acutely and severely ill, and the rapid trajectory of their decline typically drives them to proactively seek medical care. sHLH can present in patients at any age, but typically occurs in older children, adolescents and adults. Patients presenting at tertiary medical centers typically receive urgent medical intervention, including laboratory workup, supportive care, and empiric treatment. The typical progression from an underlying trigger through clinical presentation, evaluation, and treatment is illustrated below.
114
Table of Contents
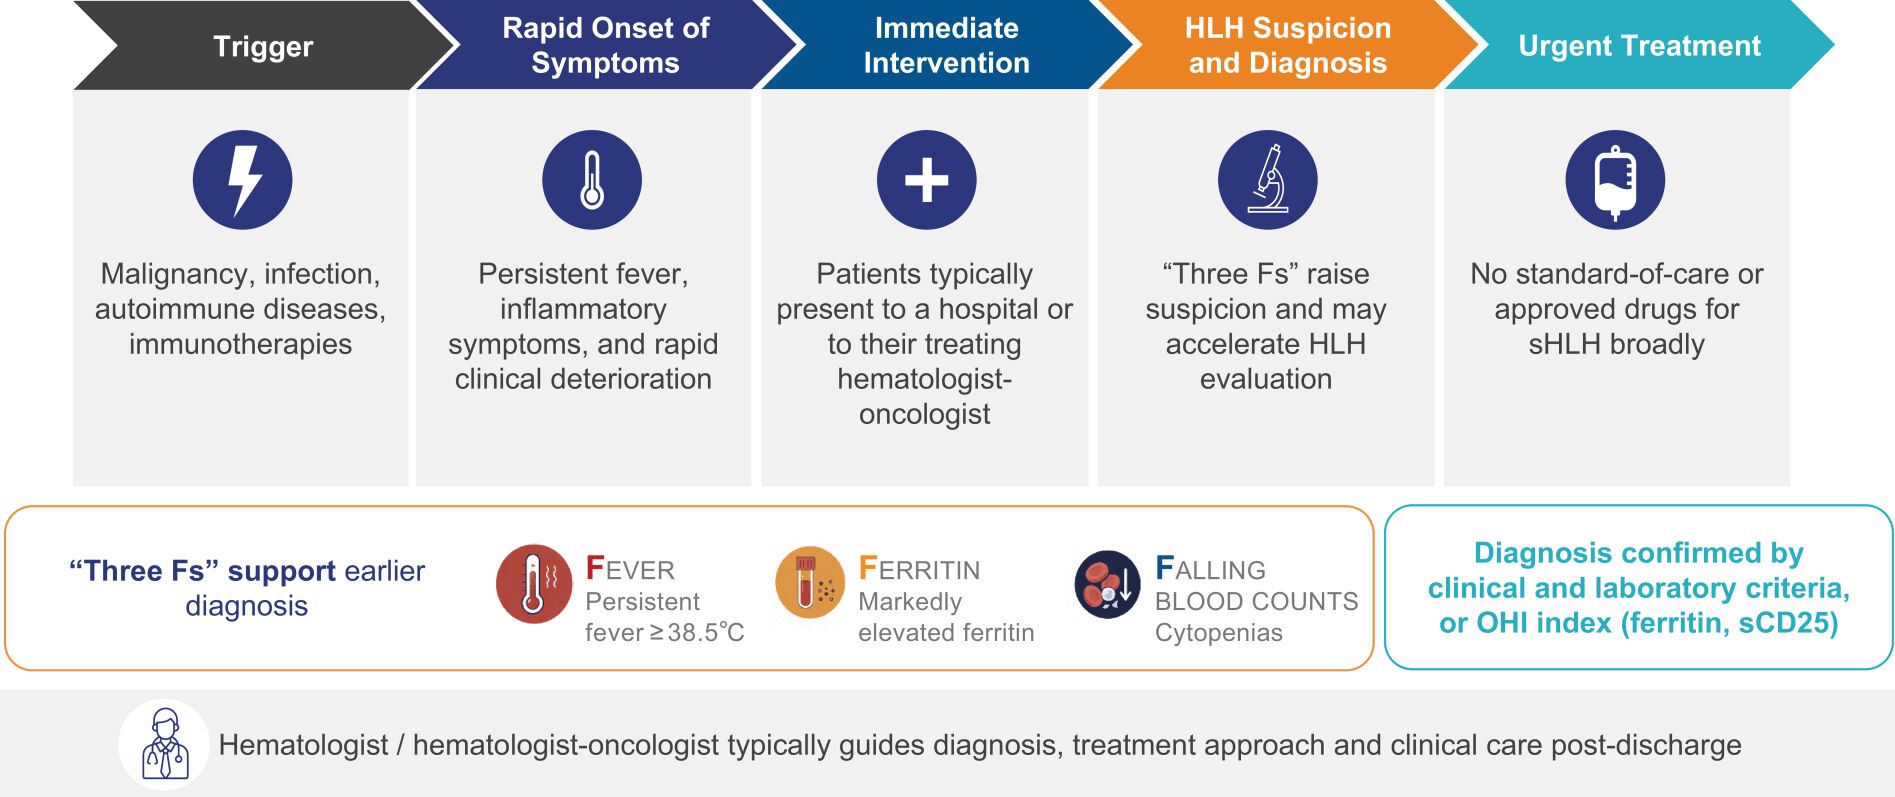
Based on the pathophysiology and presentation of sHLH, hematologist-oncologists typically direct diagnosis and treatment. Clinical suspicion and treatment of sHLH is driven by a syndrome of common signs and symptoms. A widely used mnemonic referred to as the “Three Fs” has emerged to aid recognition of the disease:
| 1. | fever that is high and unrelenting; |
| 2. | ferritin that is markedly elevated above the normal range; and |
| 3. | falling blood cell counts (cytopenias), associated with hemophagocytosis. |
Additional features frequently associated with sHLH and contributing to diagnosis include enlarged spleen and liver (splenomegaly and hepatomegaly, respectively) and elevated soluble CD25 (sCD25), a marker of T cell activation.
Other diagnostic frameworks also include the HLH-2004 criteria, originally developed for pHLH but sometimes applied across HLH subtypes, under which patients meeting at least five of eight specified clinical and laboratory features are classified as having HLH. More recently, the Optimized HLH Inflammatory (OHI) index, based on combined elevations of sCD25 and ferritin, has been validated in adults with mHLH and is gaining adoption in clinical practice. The Three Fs, HLH-2004 criteria and the OHI index are presented in the figure below:
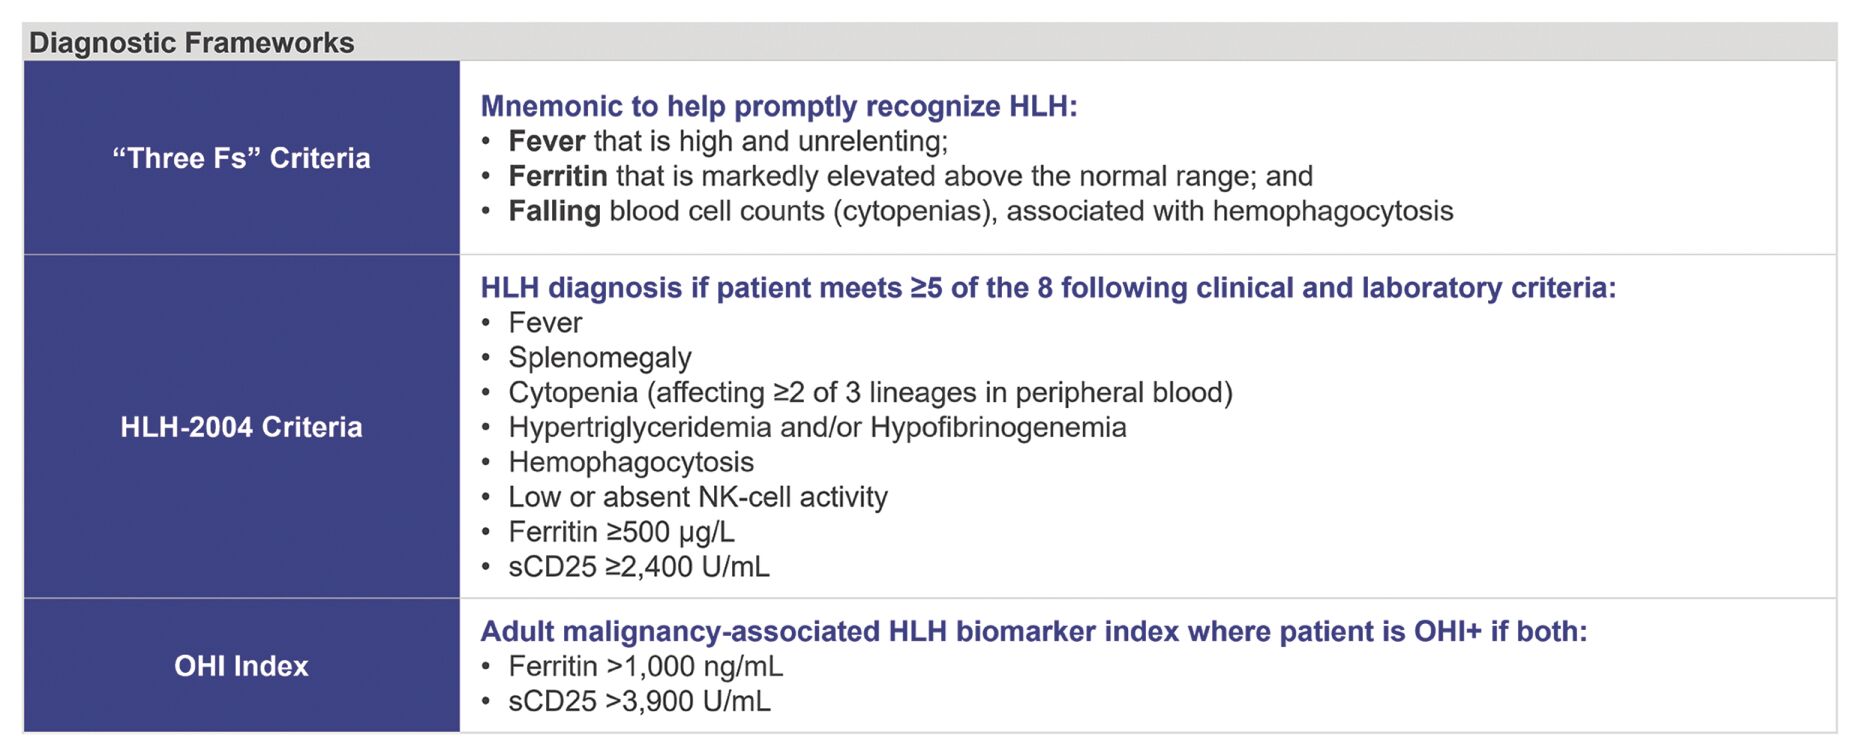
sHLH Epidemiology
Based on a primary claims data analysis using HLH-associated ICD codes, approximately 12,600 patients were diagnosed with sHLH in the United States in 2025, as shown in the chart below. Notably, the diagnosed population increased at an estimated compound annual growth rate of 11.5% from 2021 through 2025. Assuming a similar annual growth rate, this would extrapolate to a diagnosed incidence of approximately 17,500 in 2028. Moreover, we believe the diagnosed incidence of sHLH may meaningfully understate the true burden of disease because it does
115
Table of Contents
not account for misdiagnosed or undiagnosed patients, or for patients treated without assignment of an HLH-associated ICD code. Consistent with historical trends in rare diseases (such as thyroid eye disease, generalized myasthenia gravis, and hemophilia), we believe the availability of a disease-modifying therapy could meaningfully expand the patient population, as diagnosis and treatment rates increase with the availability of effective options.
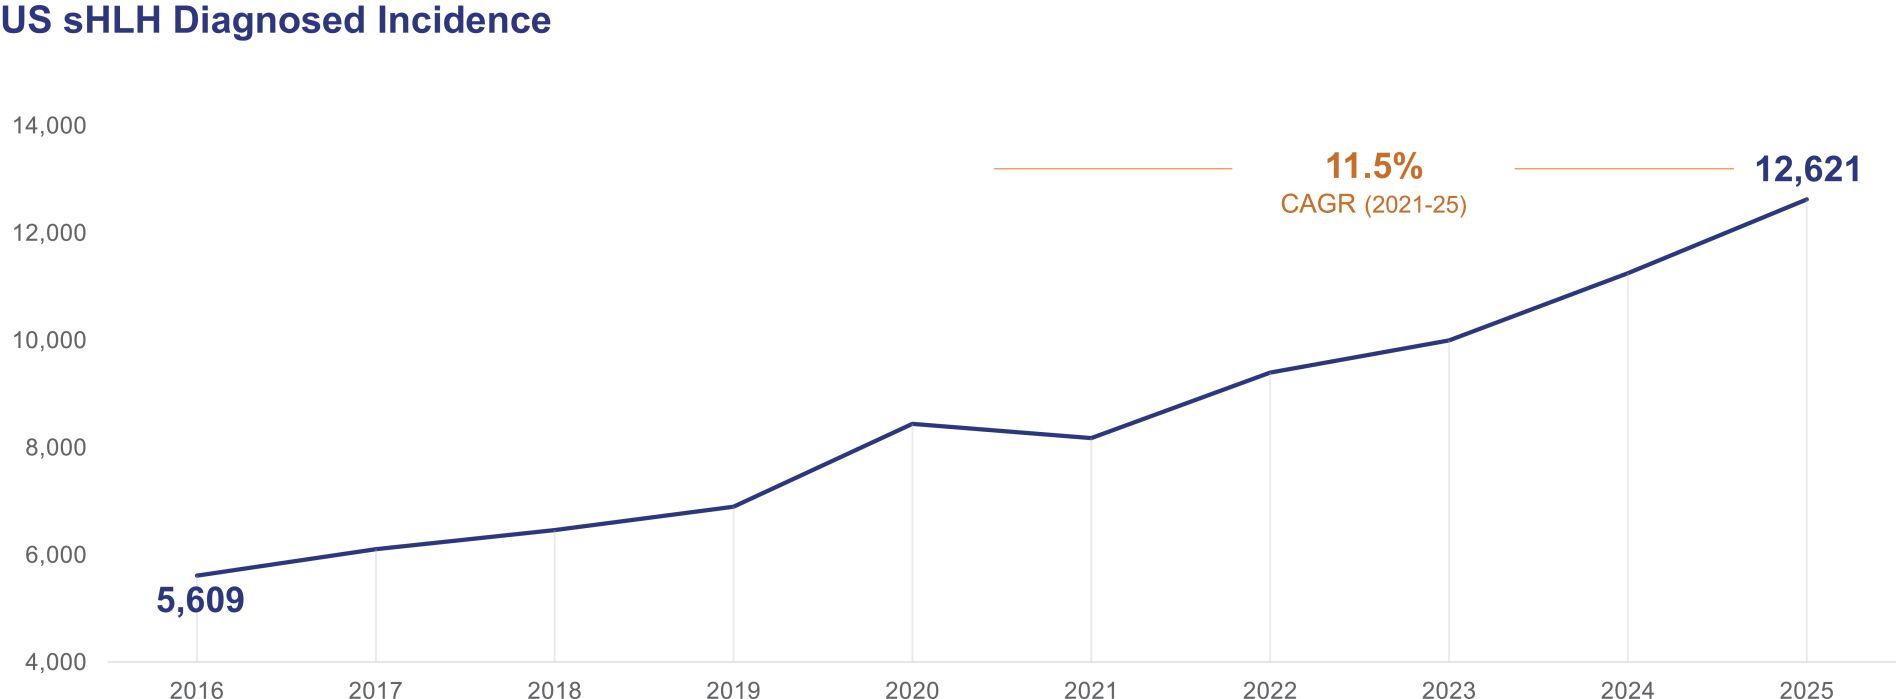
We believe several factors are driving this growth trend, including:
| ∎ | Increasing awareness and recognition of sHLH. Clinical familiarity with sHLH has increased over the past two decades, with the COVID-19 pandemic in 2020 representing an important inflection point for broader awareness within the medical community. This has contributed to improved recognition and diagnosis of sHLH across all triggers. For example, from 2021 through 2025, growth in diagnosed mHLH associated with hematologic malignancies significantly exceeded growth in the underlying hematologic malignancies themselves. |
| ∎ | Increasing incidence of underlying triggering conditions. The underlying conditions associated with sHLH, including hematologic malignancies, autoimmune diseases, and immune-activating therapies, continue to increase in incidence. In particular, expanding utilization of CAR-T therapies and bispecific TCEs is contributing to growth in the IEC-HS population. |
| ∎ | Simpler and more expedient diagnostic approaches. Increasing adoption of simpler and more practical diagnostic frameworks, including the “Three Fs” heuristic and the OHI criteria, is improving the speed and consistency of sHLH recognition and diagnosis. |
Limited Treatment Landscape: High Early Mortality Has Not Significantly Improved in Two Decades
sHLH remains a severe and rapidly progressive disease with limited treatment options and no established SOC. Despite advances in supportive care and adjacent therapeutic areas, mortality outcomes in sHLH have not significantly improved over the past two decades.
There are currently no approved therapies for the treatment of sHLH broadly. Beyond corticosteroids, such as dexamethasone, which are commonly used as background therapy, clinicians rely on a heterogeneous combination of off-label agents, including etoposide (cytotoxic chemotherapy), alemtuzumab (an anti-CD52 monoclonal antibody associated with broad immunosuppression), ruxolitinib (a JAK1/2 inhibitor), tocilizumab (an anti-IL-6 monoclonal antibody), and anakinra (an IL-1 receptor antagonist). Emapalumab (an anti-interferon-g monoclonal antibody) is FDA-approved only for refractory or recurrent pHLH and MAS associated with Still’s disease.
We believe the aforementioned therapies are limited either by broad immunosuppression or by targeting individual downstream cytokine or signaling pathways without directly addressing the activated myeloid cells and T cells that drive the inflammatory cascade in sHLH. The current patient journey, which typically involves escalating intervention without resolution and associated high early mortality, reflects these limitations.
Despite the use of these therapies, which are not specifically indicated for sHLH, many patients experience progressive multiorgan dysfunction and severe morbidity, including respiratory and hepatic failure, coagulopathy (the inability of blood to clot), and hemodynamic collapse (a life-threatening failure of the cardiovascular system), frequently resulting in escalation to ICU care and early mortality. As depicted in the image below, in-hospital
116
Table of Contents
mortality remained stable from 2007 to 2019 despite advances in supportive care and off-label use of steroids, chemotherapy, and biologic therapies.
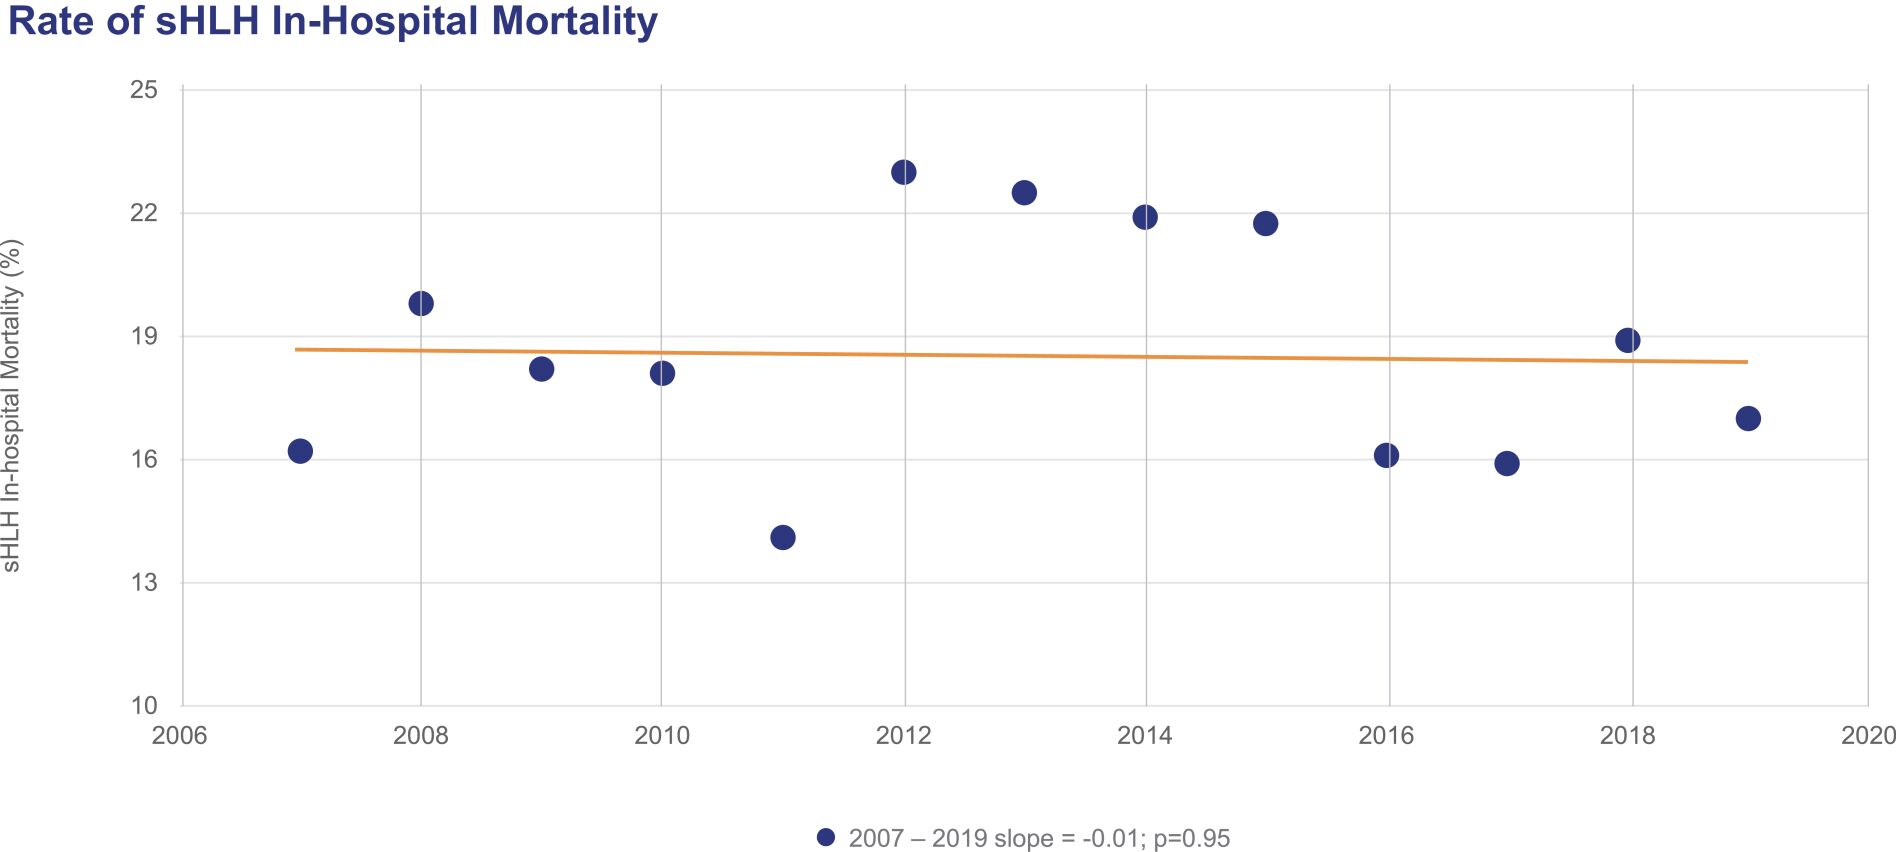
As observed across multiple studies, sHLH overall survival is severely limited, with most deaths occurring within two months of diagnosis. Among patients with mHLH, the most common and severe subtype in adults, 2-month OS with available therapies has been reported to be approximately 50%, as shown in the Kaplan Meier survival curve below. Additional publications from MD Anderson Cancer Center, Memorial Sloan Kettering Cancer Center, and Mayo Clinic also reported similar survival curves and an overall survival rate of approximately 50% at 2 months for mHLH patients.
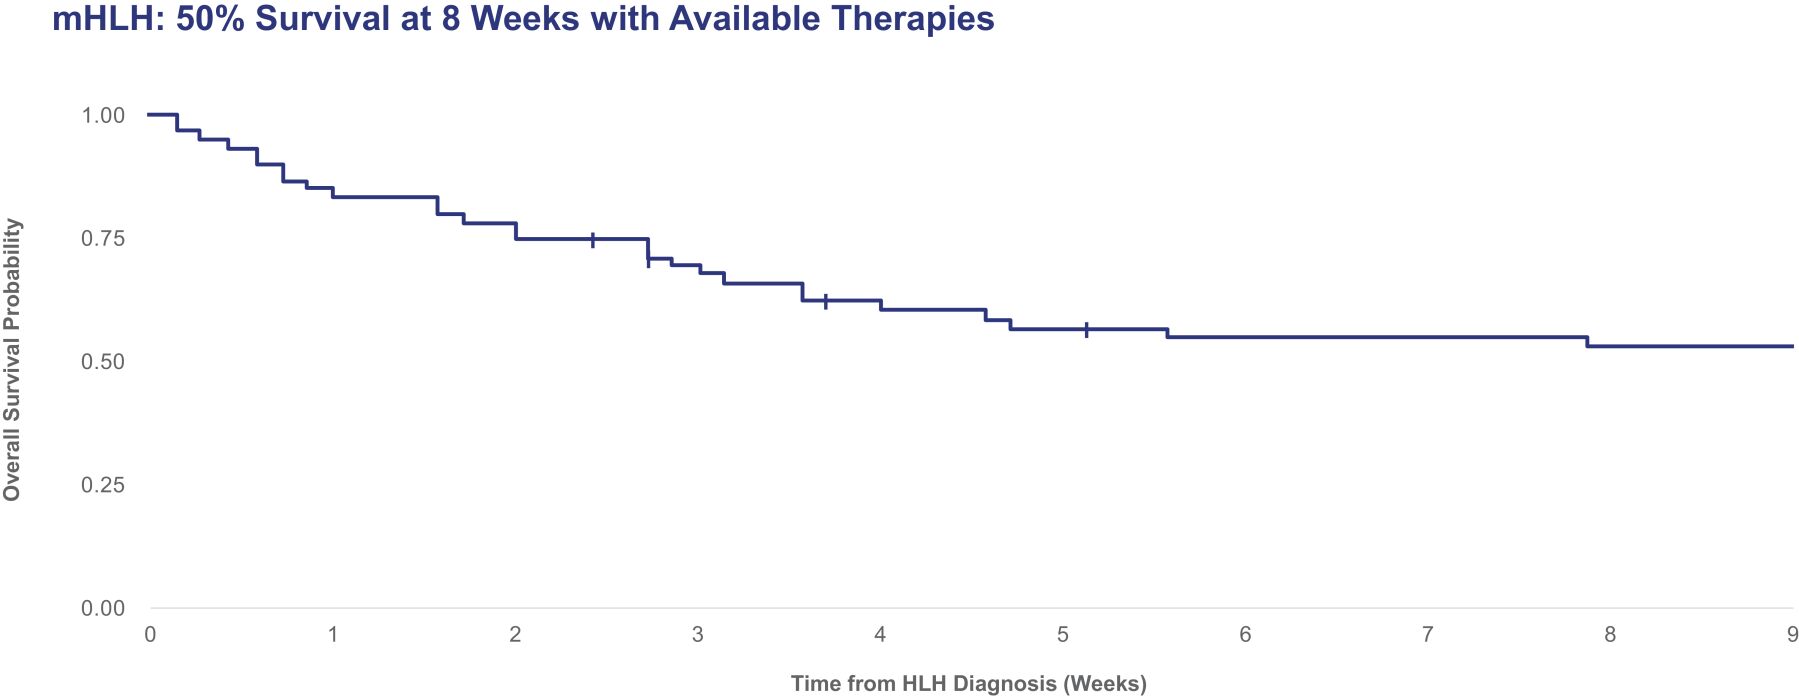
Source: mHLH patient level data at Vanderbilt University Medical Center (N=59, Broome, 2025); additional publications from MD Anderson, Memorial Sloan Kettering, and Mayo Clinic report similar survival curves and a consistent overall survival rate of ~50% at 8 weeks for mHLH patients (Long, 2025; Lee, 2021; Janda, 2025)
We believe the absence of an approved therapy for sHLH broadly, the lack of an established SOC, and persistently high early mortality underscore the critical need for more effective treatment approaches.
Our Solution: Ipsoprubart, a Pan-SIRP Antibody Targeting Myeloid Cells and T Cells
A Novel Mechanism of Action Targeting Pathogenic Immune Cells for sHLH
Ipsoprubart is an investigational, novel fully human IgG1 monoclonal antibody that binds to SIRPa and SIRPß1 on macrophages and SIRPg on T cells, immune cells that are principally responsible for driving the cytokine storm in severe hyperinflammatory conditions, including sHLH. Ipsoprubart is engineered with modifications in the fragment-crystallizable (Fc) region and is designed to bind to SIRP proteins via its complementarity-determining region, to enable both ADCC and ADCP by which antibodies eliminate target cells.
117
Table of Contents
Ipsoprubart is designed to act at the disease source by engaging Fc-receptor-bearing effector cells to rapidly eliminate the pathogenic myeloid cells and T cells that initiate and amplify the cytokine storm. Cytokine-directed inhibitors, by contrast, block only individual effectors of immune dysregulation. At the other end of the spectrum, chemotherapy agents are highly myelosuppressive in a non-specific manner, often leading to significant toxicities, including prolonged cytopenias. Because SIRP expression is elevated on activated pathogenic T cells and macrophages, ipsoprubart is designed to preferentially deplete these SIRP-high expressing cells while sparing resting and recovering immune cell populations. We believe ipsoprubart’s selectivity could translate to important safety and efficacy attributes for the treatment of inflammatory conditions. Therefore, ipsoprubart may offer a differentiated approach compared to agents currently used to treat sHLH.
Nonclinical studies demonstrated that ipsoprubart did not inhibit CD47 binding to SIRPa, suggesting preservation of the immune-surveillance signal that prior CD47-directed approaches have disrupted. Importantly, ipsoprubart’s activity has been shown to be driven by Fc-mediated depletion of pathogenic SIRP-expressing myeloid cells and T cells.
Given its novel mechanism of action, we believe ipsoprubart can rapidly disrupt the cytokine storm at its source, offering the potential to become the first therapy for sHLH broadly, irrespective of the underlying trigger or cytokine expression pattern.
Ipsoprubart is designed for subcutaneous (SC) or intravenous (IV) administration in either outpatient or inpatient settings. For the treatment of sHLH, patients in our trials generally start ipsoprubart in the inpatient setting for priming and loading doses. Once patients are stabilized and discharged from the hospital, they continue treatment in the outpatient setting for most of their treatment course.
Nonclinical Pharmacology and Toxicology
In nonclinical studies, ipsoprubart demonstrated comparable binding to human and NHP SIRPa, SIRPß1, and SIRPg isoforms, with binding affinities within three-fold across paired isoforms, supporting the translational relevance of the NHP model for IND-enabling toxicology.
Ipsoprubart induced ADCC and ADCP of primary human and NHP monocytes and T cells in vitro, with EC90 values of 0.1 µg/mL or less. IV administration of ipsoprubart to NHPs demonstrated dose-proportional exposure, as measured by area under the concentration–time curve (AUC) and maximum observed plasma concentration (Cmax), and duration of plasma ipsoprubart concentrations. SC administration of ipsoprubart displayed substantial bioavailability. Administration of ipsoprubart to NHPs resulted in rapid and reversible depletion of circulating monocytes and lymphocytes, with a dose-dependent PK-PD effect at target dose ranges.
At plasma concentrations of ipsoprubart above approximately 0.1 µg/mL, depletion of target cell populations was sustained, and once plasma concentrations fell below this threshold, target cell counts returned to baseline. We believe this reversibility may represent a key safety differentiator compared with therapies such as etoposide and alemtuzumab, as etoposide can cause broad myelosuppression through effects on proliferating hematopoietic progenitors, while alemtuzumab has been associated with profound and prolonged lymphodepletion that may delay immune reconstitution. Ipsoprubart was well tolerated in both 28-day and 6-month GLP repeat-dose toxicology studies in NHPs.
Robust Clinical Program and Regulatory Designations in sHLH
We have completed a Phase 1 clinical program for ipsoprubart comprising two trials:
| ∎ | A Phase 1 randomized, double-blind, placebo-controlled, multicenter trial in 94 healthy adult volunteers, evaluating the safety, tolerability, PK, and PD of ipsoprubart following single and multiple doses administered IV and/or SC. |
| ∎ | A Phase 1b open-label, single-arm, multicenter trial in 22 patients with sHLH, evaluating the safety, efficacy, PK, and PD of ipsoprubart and identifying a dosing regimen for further clinical evaluation. |
Following completion of the Phase 1 program, we received FDA feedback on a registrational development program for ipsoprubart in sHLH. We have initiated two trials intended to support potential approval of ipsoprubart. This includes SURPASS, our Phase 2/3 registrational trial of ipsoprubart in newly diagnosed, treatment-naïve patients with sHLH, and COMPASS, a natural history study from which protocol-defined data collection and systematic clinical adjudication will support the selection of the external control comparator for SURPASS.
118
Table of Contents
In 2025, ipsoprubart received BTD from the FDA and PRIME designation from the EMA. Both designations are for the treatment of sHLH broadly, which we believe reflects the acknowledgment by the regulatory authorities of the high unmet need in sHLH and that our data demonstrate the potential for ipsoprubart to provide substantial improvements over currently available therapy on one or more clinically meaningful endpoints, including overall survival.
Our Completed Phase 1 Clinical Trial in Healthy Volunteers
We have completed a Phase 1, randomized, double-blind, placebo-controlled trial of ipsoprubart in healthy adult volunteers. The trial was designed to evaluate the safety, tolerability, PK, and PD of single and multiple ascending doses of ipsoprubart administered IV and/or SC, but it was not powered for statistical significance. A total of 94 participants received ipsoprubart or placebo across two parts.
In Part 1, the single-ascending-dose portion of the trial, 50 participants received single doses of ipsoprubart administered IV (0.001 to 0.01 mg/kg) or SC (0.02 to 0.5 mg/kg), and 17 received placebo. In Part 2, the multiple-dose portion of the trial, 18 participants received multiple SC doses of ipsoprubart at cumulative doses up to 1.2 mg/kg, and 9 received placebo. The trial was conducted in Europe, and all 94 participants completed the trial.
Ipsoprubart produced substantial transient reductions in monocyte and lymphocyte counts beginning at the 0.1 mg/kg dose level, with effects greater than those observed in placebo-treated participants at all dose levels at or above 0.1 mg/kg. In the multi-dose cohorts, repeated dosing demonstrated that these effects occurred with each dose, supporting the intended PD activity of ipsoprubart in humans.
PK modeling based on plasma concentration data from this trial supports a twice-weekly dosing regimen of 0.5 mg/kg, which we predict will maintain steady-state plasma concentrations above the PD threshold of 0.1 µg/mL with minimal drug accumulation.
Our Completed Phase 1b Trial in sHLH
We completed a Phase 1b open-label, single-arm, multicenter trial in 22 patients with sHLH, evaluating the safety, efficacy, PK, and PD of ipsoprubart and identifying a dose regimen for further clinical evaluation. The trial was not powered for statistical significance. The trial was commenced in May 2022 with the last known follow-up with a living patient in March 2025, with no further follow-up anticipated. We believe this completed trial demonstrated compelling clinical proof-of-concept for ipsoprubart, reflecting its potential in treating sHLH broadly, regardless of the trigger. Ipsoprubart was evaluated across three sequential cohorts that informed both the patient population and the dosing regimen advanced into our ongoing Phase 2/3 program.
Trial Design and Population Evolution
Across the Phase 1b trial, 22 patients with sHLH received ipsoprubart across three cohorts. Cohort 1 evaluated daily dose escalation of ipsoprubart from 0.1 to 3.0 mg/kg IV. Cohort 2 evaluated a priming dose (0.1 mg/kg IV) and three daily loading doses (0.3 mg/kg IV), followed by 1.0 mg/kg weekly maintenance dosing (SC or IV). Cohort 3 evaluated the same priming and loading regimen as Cohort 2, followed by 0.5 mg/kg administered twice weekly. These cohorts informed the dosing regimen selected for further ipsoprubart clinical development in sHLH, consisting of a 0.1 mg/kg priming dose, followed by three 0.3 mg/kg daily loading doses, followed by twice-weekly 0.5 mg/kg maintenance dosing, administered SC or IV over a 12-week treatment course, with dexamethasone as background therapy. In our Phase 1b trial, the 0.3 mg/kg loading doses rapidly achieved clinically meaningful drug concentrations, and 0.5 mg/kg maintenance dosing sustained levels above the target threshold of 0.1 µg/mL throughout the treatment course.
Cohorts 1 and 2 enrolled a total of 12 patients who were either treatment-naïve or relapsed/refractory to prior HLH-directed therapy. Across these cohorts, the ORR was 75% (9/12) by Week 4. Response was defined as either CR, modified CR (mCR), partial response (PR), or HLH improvement (HI), with no progression of other aspects of HLH disease pathology. A CR is defined as resolution of all clinically relevant manifestations of HLH, including fever, splenomegaly, cytopenias, hyperferritinemia, coagulopathy, neurologic abnormalities, and cerebrospinal fluid abnormalities, as applicable, and absence of sustained worsening in sCD25 levels. An mCR is equivalent to a CR except that normalization of splenomegaly and cytopenias is not required. A PR is achieved when at least three HLH abnormalities meet CR criteria and there is no progression of other aspects of HLH disease pathology. HI is defined as a >50% improvement from baseline in at least three HLH clinical or laboratory abnormalities.
Data from Cohorts 1 and 2 demonstrated a marked divergence in survival between the two populations, where treatment-naïve patients achieved a median OS (mOS) of 152 days (approximately 22 weeks) and the relapsed/
119
Table of Contents
refractory patients, who received prior therapies, typically presented with more advanced multiorgan failure from prolonged sHLH activity, had a mOS of 24 days. Based on these findings, the independent Data Monitoring Committee (DMC) recommended that the trial be redirected to focus on frontline treatment with ipsoprubart.
Cohort 3 enrolled 10 patients in the frontline setting who were either treatment-naïve or early refractory, defined as having received less than one week of prior HLH-directed therapy without sufficient clinical or laboratory response. The schematics of our Phase 1b trial design are shown in the figure below:
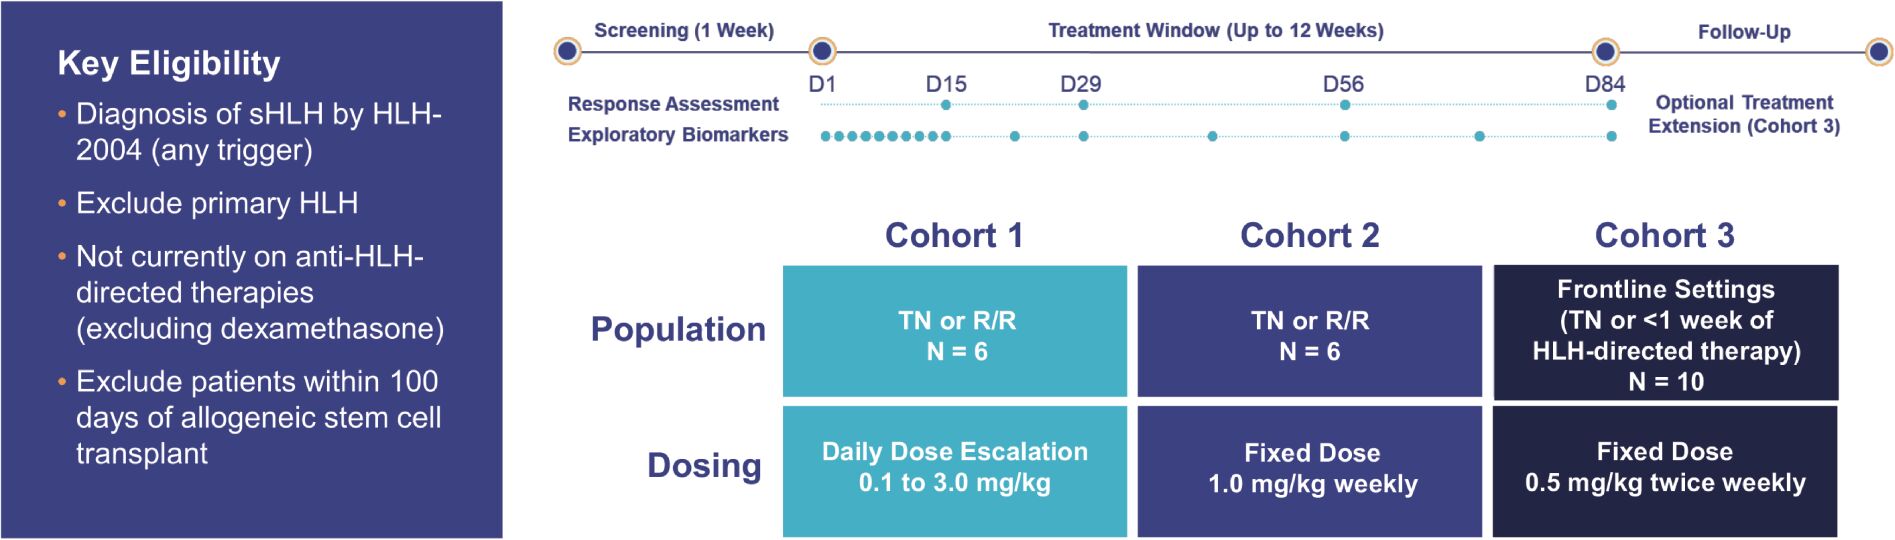
TN: Treatment Naïve; R/R: Relapsed / Refractory
Baseline Characteristics of Frontline mHLH Population
We assessed clinical activity of ipsoprubart in mHLH, which represents the majority of patients enrolled in our Phase 1b trial and accounts for approximately 45% of all sHLH cases. The Phase 1b trial was broad at the outset and enrolled both treatment-naïve and relapsed/refractory sHLH patients across cohorts to characterize safety, PK/PD, dose regimen, and early clinical activity. The clinical activity analysis was focused on the 12 frontline mHLH patients because this is the largest subset and represented the most clinically interpretable population and most directly informed the primary efficacy population for the registrational trial.
The baseline characteristics of the 12 frontline mHLH patients were generally representative of, and in several respects more severe than, the broader mHLH population.
| ∎ | 10 of the 12 patients had lymphoma as their underlying malignancy trigger. |
| ∎ | Of the 10, 7 had T cell lymphoma, a subtype associated with poor outcomes independent of sHLH. |
| ∎ | 5 of the 12 patients had relapsed/refractory underlying malignancy at enrollment, an independent adverse prognostic factor. |
More details on baseline characteristics of the frontline mHLH patient population in the Phase 1b trial are summarized in the table below.
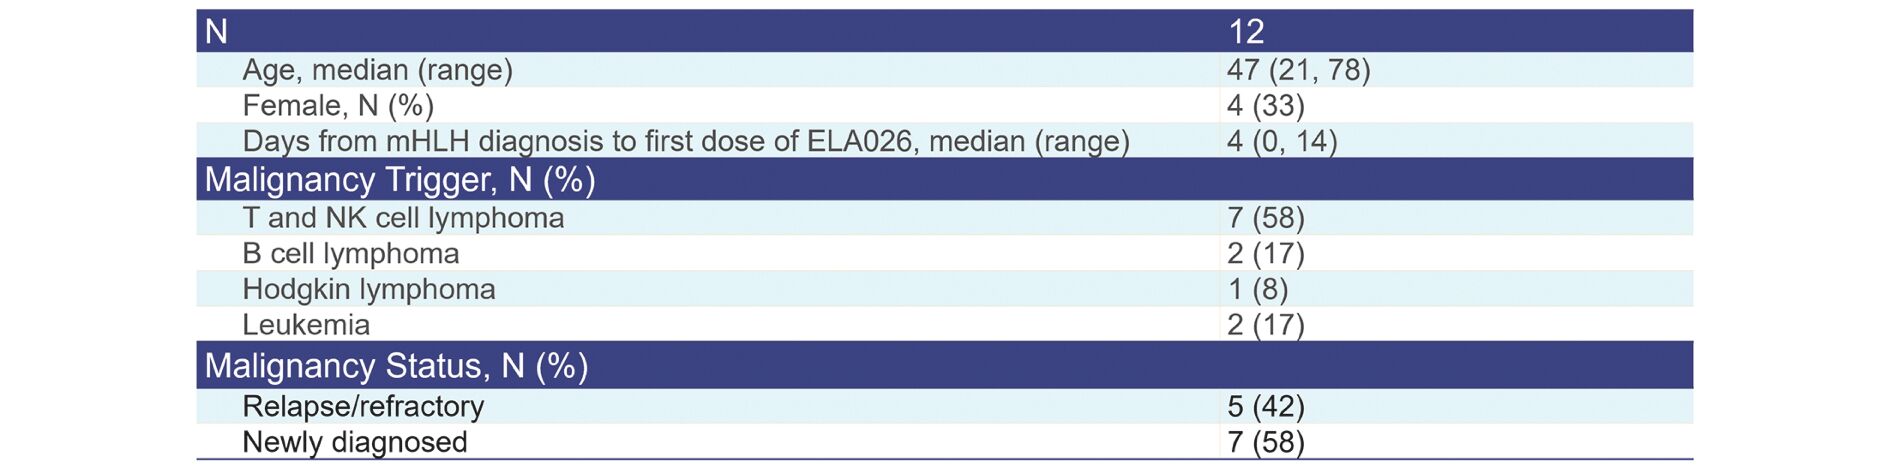
120
Table of Contents
100% Overall Survival at Week 8 in Frontline mHLH Population
Frontline ipsoprubart treatment produced a 100% OS rate at 8 weeks, from the time of diagnosis, as shown in the Kaplan Meier survival curve below. Notably, deaths occurring beyond 8 weeks were commonly attributed to the underlying refractory malignancy, and not to HLH.
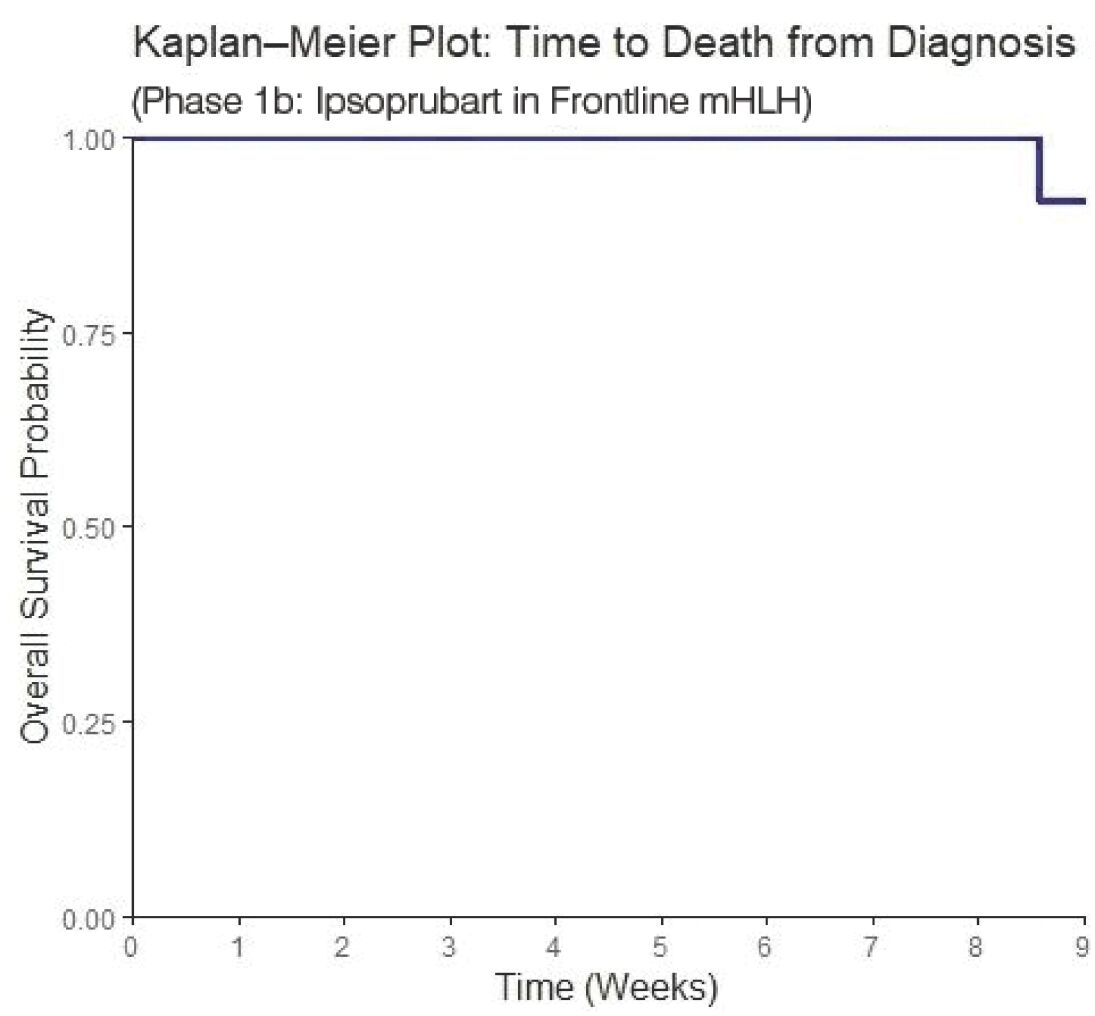
100% Overall Response and 100% Hospital Discharge in Frontline mHLH Population
Frontline ipsoprubart treatment produced a 100% ORR by the Week 4 assessment in mHLH patients, as assessed by modified HLH-2004 response criteria. This reflects improvement or resolution of the clinical and laboratory features associated with the initial sHLH diagnosis. Of the 12 evaluable patients at the Week 4 assessment, 10 achieved a PR and two achieved an mCR. Notably, the modified HLH-2004 response criteria were developed for use in pHLH, rather than mHLH. Consequently, a CR may not have been achievable in some patients in our trial because CR criteria requires normalization of variables, such as ferritin levels, which can remain affected by the underlying malignancy or ongoing chemotherapy, even when HLH disease activity is effectively controlled.
Ipsoprubart also produced a 100% hospital discharge rate amongst 11 frontline mHLH patients evaluable for discharge (one patient withdrew prior to discharge assessment), which we view as a meaningful endpoint in terms of both clinical and health economic outcomes. Surviving the initial hospitalization reflects stabilization of the hyperinflammation in sHLH and is a prerequisite for deriving longer-term benefit. It enables completion of a diagnostic evaluation, informed treatment decision-making with patients and their families, and initiation of definitive anti-cancer therapy. Transition from inpatient to outpatient care may also reduce healthcare resource utilization and costs.
Rapid Improvement of Pharmacodynamic Biomarkers in Frontline mHLH Population
PD data support our belief that ipsoprubart acts by depleting pathogenic immune cells in sHLH and has the potential to rapidly normalize inflammatory disease activity. As shown in the figure below, we observed deep reductions across three biomarkers central to sHLH pathophysiology: a 61% median reduction in ferritin; a 69% median reduction in sCD25; and an 83% median reduction in C-reactive protein (CRP). Ferritin and sCD25 are markers of macrophage and T cell activation, respectively, and represent two of the eight HLH-2004 diagnostic criteria, and CRP is a marker of systemic inflammation. These biomarker reductions were rapid following treatment initiation. In seven mHLH patients with chemotherapy-free treatment windows, ipsoprubart produced average reductions of approximately 40% in ferritin and 50% in CRP by Day 6 of treatment.
121
Table of Contents
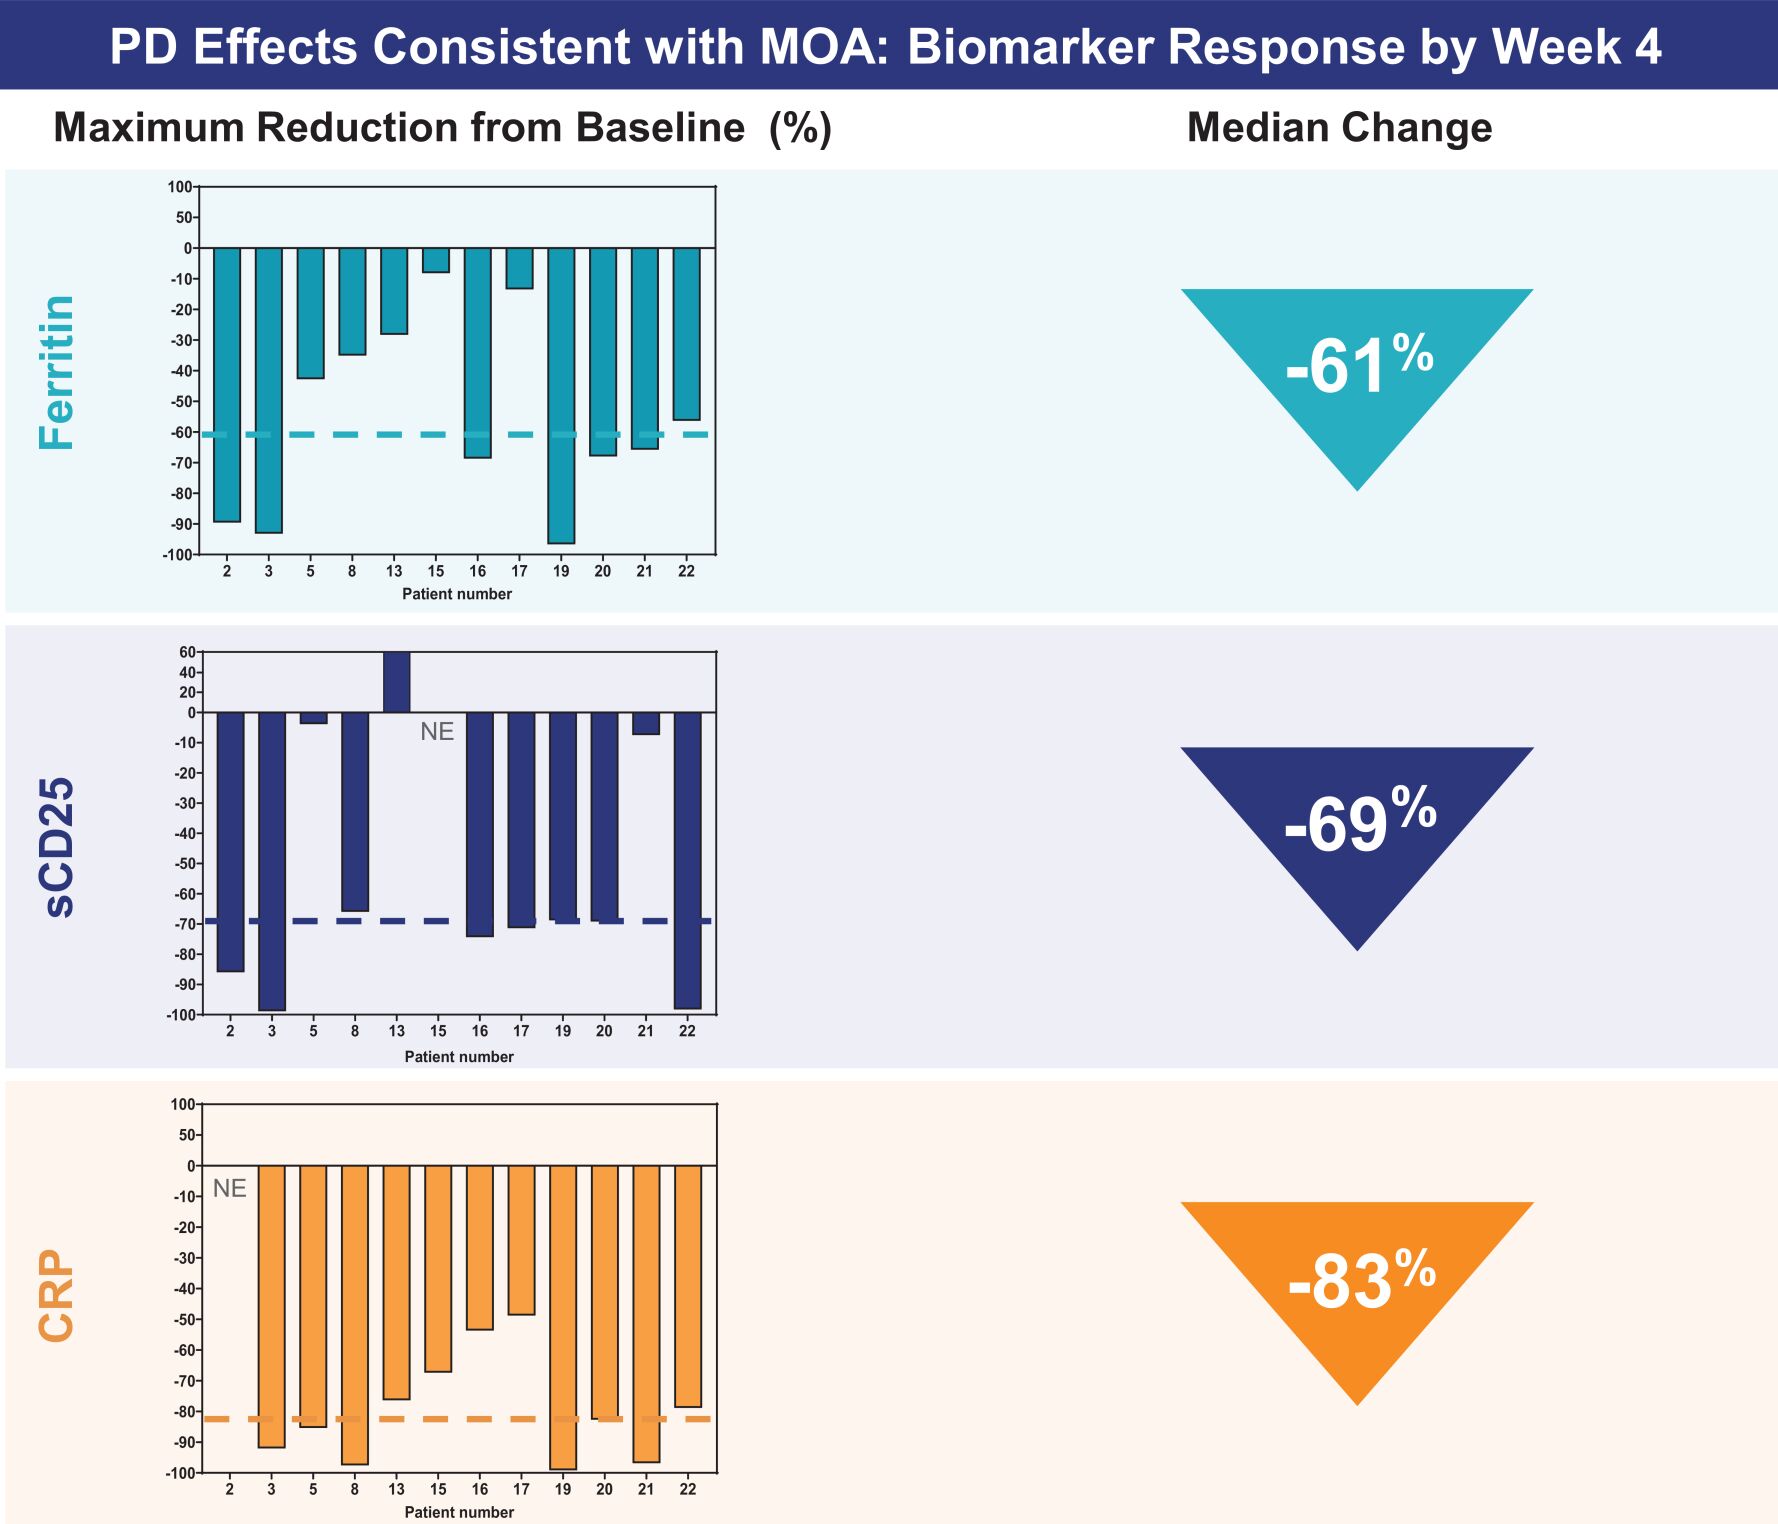
In addition, we observed reductions in HLH-associated inflammatory cytokines of patients whose samples were available for analysis, including interferon-g, C-X-C motif chemokine ligand (CXCL10), IL-8, IL-10, IL-18, and CXCL9, that correlated with clinical response. Cytokine data are presented as fold change from baseline for mHLH patients treated with ipsoprubart in the frontline setting. Values were normalized to baseline and are shown as fold increases (red) or fold reductions (green) relative to the baseline sample at each study day.
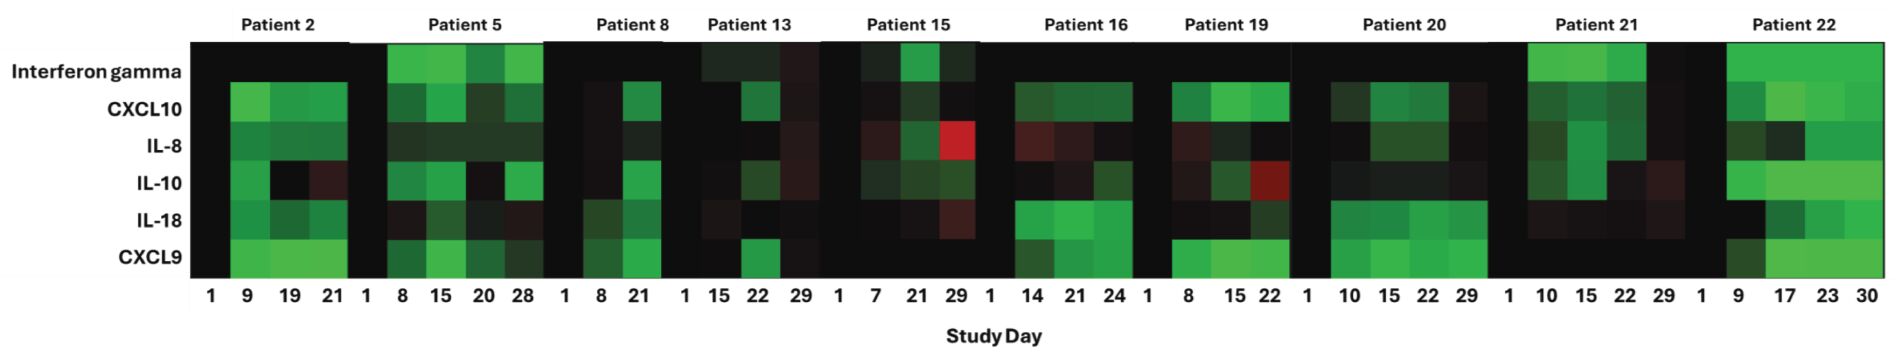
Two Case Studies Highlight Ipsoprubart Monotherapy Activity Across sHLH, Including Non-Malignancy Associated Disease
Complete Response in EBV-triggered sHLH with Frontline Ipsoprubart Monotherapy
Our Phase 1b trial included a patient case of iHLH triggered by Epstein-Barr virus (EBV), which we believe supports the potential applicability of ipsoprubart across sHLH triggers.
A 20-year-old female patient presented with acute EBV infection accompanied by associated neurological symptoms, liver function abnormalities, and severe neutropenia. She was admitted to the hospital, diagnosed with sHLH by a hematologist-oncologist, and was treatment-naïve for sHLH at the time of trial entry.
122
Table of Contents
The patient received the Cohort 3 dose regimen and completed a 12-week course of ipsoprubart monotherapy with background dexamethasone. She experienced rapid and clinically meaningful improvement, including resolution of neurological symptoms within days and normalization of ferritin and cytopenias by Week 2. The EBV infection resolved. She achieved an mCR by Week 2 and a confirmed CR by Week 8 as assessed by modified HLH-2004 response criteria.
Following completion of the 12-week treatment course, ipsoprubart was discontinued, and the patient remained in full remission more than one year later. Ipsoprubart was well tolerated throughout treatment, with no adverse events assessed as related to trial drug and no evidence of clinically meaningful neutropenia or thrombocytopenia.
Complete Response in mHLH Patient Treated with Frontline Ipsoprubart Monotherapy
A 26-year-old male patient presented with sHLH triggered by refractory cutaneous cytotoxic CD8+ T cell lymphoma. The patient presented with bone marrow failure, massive splenomegaly, nodal disease, and transfusion dependence. He had previously received more than five lines of anti-cancer therapy and accessed ipsoprubart through an Expanded Access Program.
The patient received the Cohort 3 dose regimen and ipsoprubart monotherapy, with no added HLH or anti-cancer therapy, for the entire 12-week treatment course. He experienced rapid and clinically meaningful improvement, with palpable spleen reduction within weeks of treatment. There were no adverse events related to ipsoprubart during treatment, and he achieved a complete HLH response by Week 4 and a complete tumor response by Week 13.
Following completion of the 12-week treatment course, ipsoprubart was discontinued. The patient achieved complete HLH and tumor responses and was alive as of the last follow-up, which was about one year later. Additional details of the tumor response are described below in the subsection titled "—Ipsoprubart: Expansion into Hematologic Malignancies."
These two patient cases illustrate the applicability of ipsoprubart across distinct sHLH triggers when administered as monotherapy for 12 weeks. Both patients achieved rapid control of HLH and complete responses. Treatment consisted of a 12-week course of ipsoprubart, administered predominantly in the outpatient setting, and was well tolerated, with no drug-related cytopenias, infections, or other clinically significant adverse events observed.
Ipsoprubart Generally Well Tolerated in the Phase 1 Program
Across our Phase 1 and Phase 1b trials, ipsoprubart was generally well tolerated in the population. Among the sHLH population in our Phase 1b trial (n=22), as shown in the table below, the most frequently reported adverse events overall were pyrexia, hyperkalemia, and hypotension, which are consistent with the underlying severity of sHLH and its triggers. The most common treatment-related adverse events in the Phase 1b trial included infusion-related reactions, neutropenia (neutrophil count decrease), and thrombocytopenia (platelet count decrease). Infusion-related reactions and their associated symptoms, such as fever, rigors, and chills, were observed primarily with the initial doses and were generally managed with standard premedication and infusion-management practice, consistent with the expected pharmacology of cell-depleting antibodies. Serious adverse events of Grade 5 in our Phase 1b trial occurred predominantly in patients with relapsed/refractory sHLH who presented with multiorgan failure or refractory underlying malignancy at baseline. No Grade 5 event was assessed by the investigator as related to ipsoprubart. We believe the overall safety data set observed in the Phase 1 and Phase 1b trials is consistent with the profile of critically ill sHLH patients.
123
Table of Contents
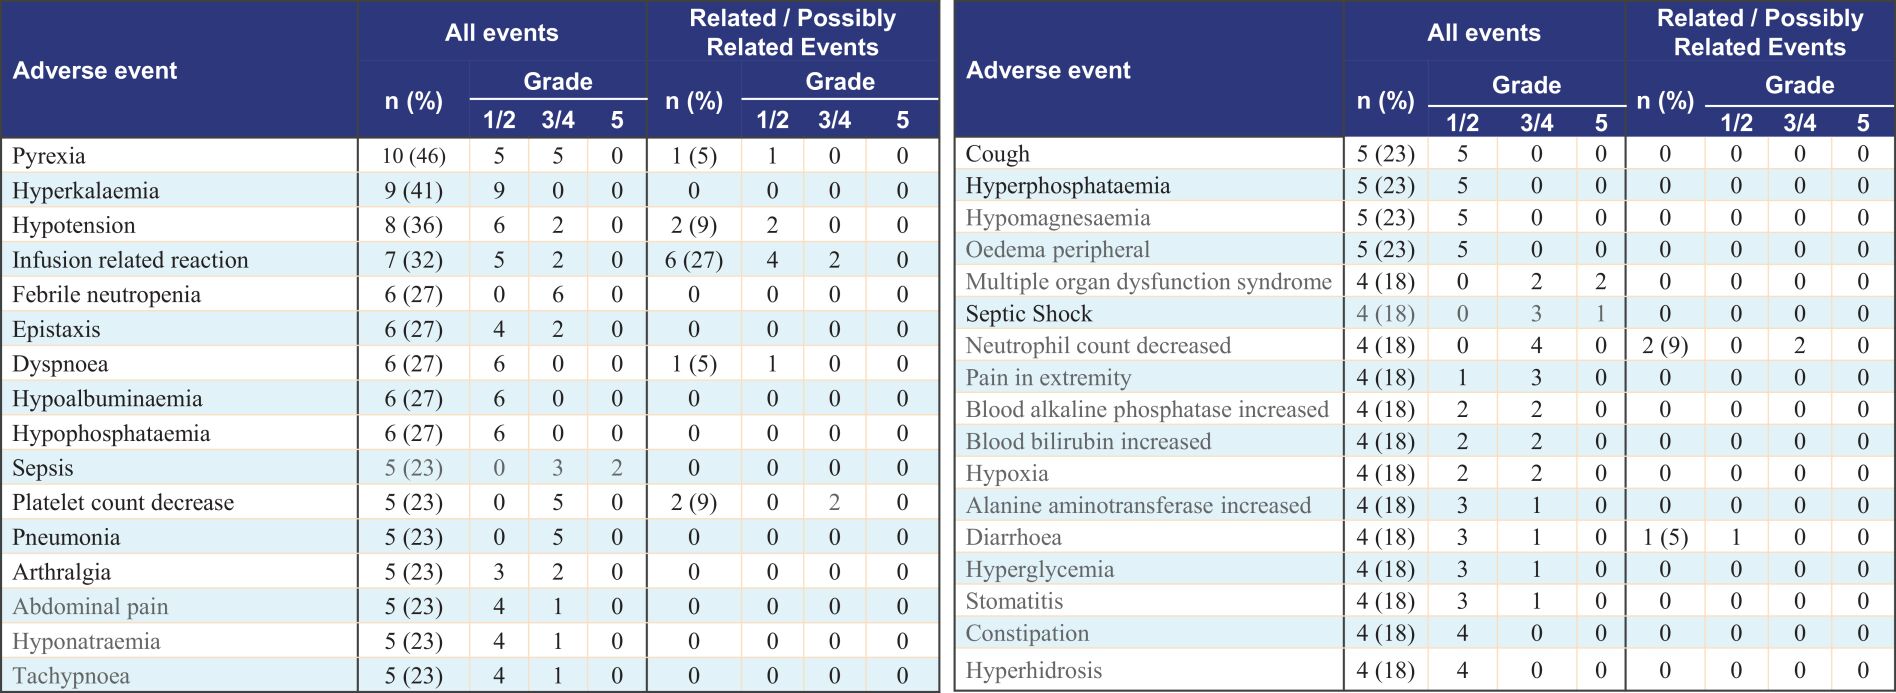
Treatment emergent adverse events occurring in greater than 15% of patients (n=22).
Global Registrational Program Designed to Support Approval of Ipsoprubart for sHLH
Following the positive results from our Phase 1b trial and feedback from the FDA on a pivotal development plan, we initiated a registrational program for ipsoprubart in sHLH consisting of: SURPASS, our global Phase 2/3 registrational trial, and COMPASS, our natural history study designed to provide an external control comparator for SURPASS.
SURPASS: Global Phase 2/3 Registrational Trial
SURPASS is a global, open-label, single-arm, multicenter, external-controlled Phase 2/3 registrational trial evaluating ipsoprubart in adult and pediatric patients with newly diagnosed, treatment-naïve sHLH enrolled into two cohorts:
| ∎ | Cohort A (primary cohort) includes patients with newly diagnosed, treatment-naïve mHLH diagnosed by HLH-2004 criteria and triggered by any malignancy. Of the up to 70 patients to be enrolled in Cohort A, we expect to enroll approximately 47 evaluable patients with lymphoma-associated HLH (LA-HLH), which will serve as the primary efficacy analysis population. LA-HLH is the largest subset of mHLH patients and represented 83% (10 of 12) of the treatment-naïve mHLH patients enrolled in our Phase 1b trial. By designating LA-HLH as the primary efficacy analysis population, we aim to reduce clinical heterogeneity and enable a focused propensity score-weighted analysis against a comparable external control population captured in COMPASS. |
| ∎ | Cohort B (exploratory cohort) includes patients across three subpopulations to generate additional safety and efficacy experience in: (1) adults aged 18 years and older with non-malignancy-associated sHLH, (2) adults aged 18 years or older with mHLH diagnosed by OHI criteria but not meeting HLH-2004 criteria; and (3) pediatric patients aged 6 to 17 years with newly diagnosed sHLH of any trigger. Enrollment in pediatric patients aged 6 to 12 years will be governed by a DMC-supervised safety lead-in cohort. Within Cohort B, we plan to enroll up to 50 patients. |
COMPASS: Natural History Study and External Control Comparator
COMPASS is a retrospective natural history study that is designed to provide an external control comparator for the SURPASS primary efficacy analysis. COMPASS is designed to characterize clinical outcomes in patients with sHLH treated with currently available therapies, which are not currently approved specifically for sHLH, under real-world conditions. The study identifies patients with HLH from electronic medical records at participating institutions, confirms the sHLH diagnosis through clinical adjudication and chart review, and applies similar eligibility criteria as the SURPASS LA-HLH primary efficacy analysis population. COMPASS is designed to provide at least 94 LA-HLH patients to serve as the external control comparator.
Primary Endpoint and Analytical Approach
The primary endpoint of SURPASS is 8-week OS from time of sHLH diagnosis in the LA-HLH population, compared against a propensity-score-weighted external control population from COMPASS. The propensity-score-weighted analytical framework governing the comparison between SURPASS and COMPASS is pre-specified in a statistical analysis plan developed in consultation with the FDA.
124
Table of Contents
The 8-week timepoint was selected as the primary endpoint because: (1) it captures the period of greatest HLH-related mortality, during which the treatment effect of ipsoprubart on the underlying immune dysregulation is most directly observable; (2) it balances the time needed to demonstrate a meaningful treatment effect against the period beyond which competing mortality events, such as malignancy progression or chemotherapy-related toxicity, become significant confounders; and (3) it is supported by the observation in our Phase 1b trial that deaths occurring beyond Week 8 were attributed to the underlying malignancy, rather than HLH. The analytical framework applies propensity score methodology to adjust for differences in key baseline covariates between the SURPASS and COMPASS patient populations.
Secondary and Exploratory Endpoints
Secondary endpoints of SURPASS include OS at 30, 45, 56 (the primary endpoint in LA-HLH and secondary endpoint in broader mHLH and sHLH populations), and 90 days; survival status at hospital discharge; best HLH disease response by Day 29 as assessed by modified HLH-2004 criteria; and incidence of treatment-emergent adverse events. Exploratory endpoints include longer-term measures of OS, tumor response, PK and PD changes over time, and other clinical outcomes.
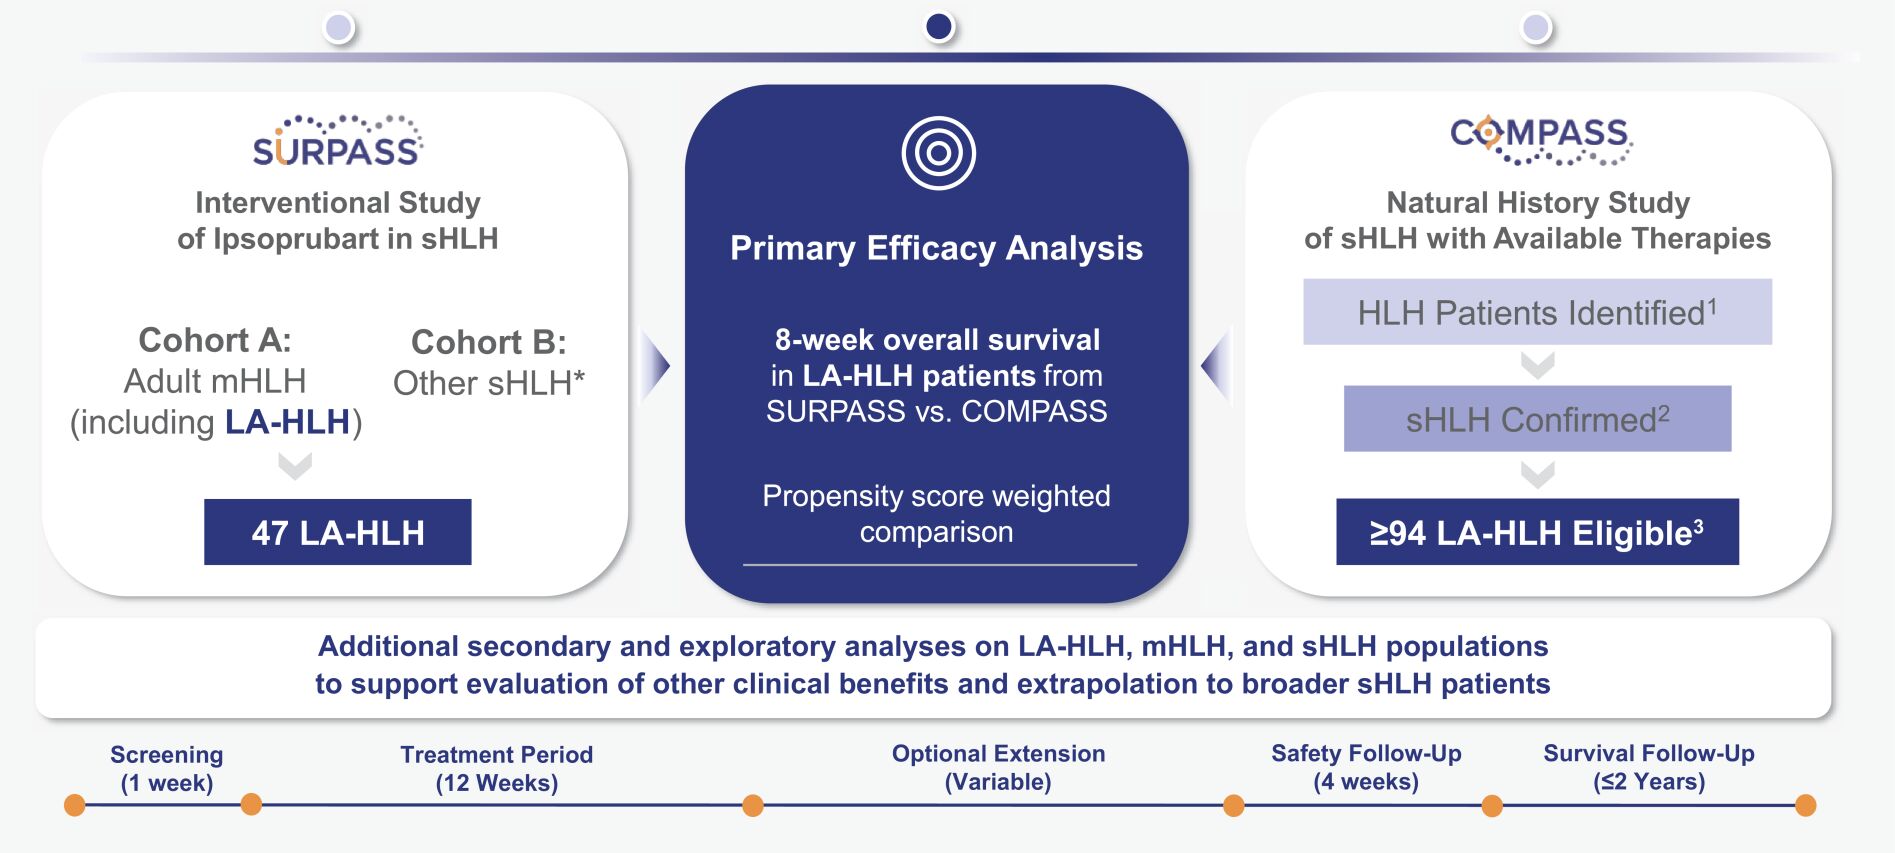
*Other types of sHLH include: a) non-mHLH subtypes in patients ≥18 years; b) mHLH in patients ≥18 years old, diagnosed by OHI+ (optimized HLH index) criteria but not meeting HLH-2004 criteria; c) any sHLH type in patients aged 6 to 17
1. HLH patients identified from EMR.
2. sHLH patients confirmed by clinical adjudication through chart review.
3. LA-HLH subjects meeting SURPASS inclusion/exclusion criteria selected for historical control.
Dosing Regimen
SURPASS patients will receive the dosing regimen used in Cohort 3 of our Phase 1b trial:
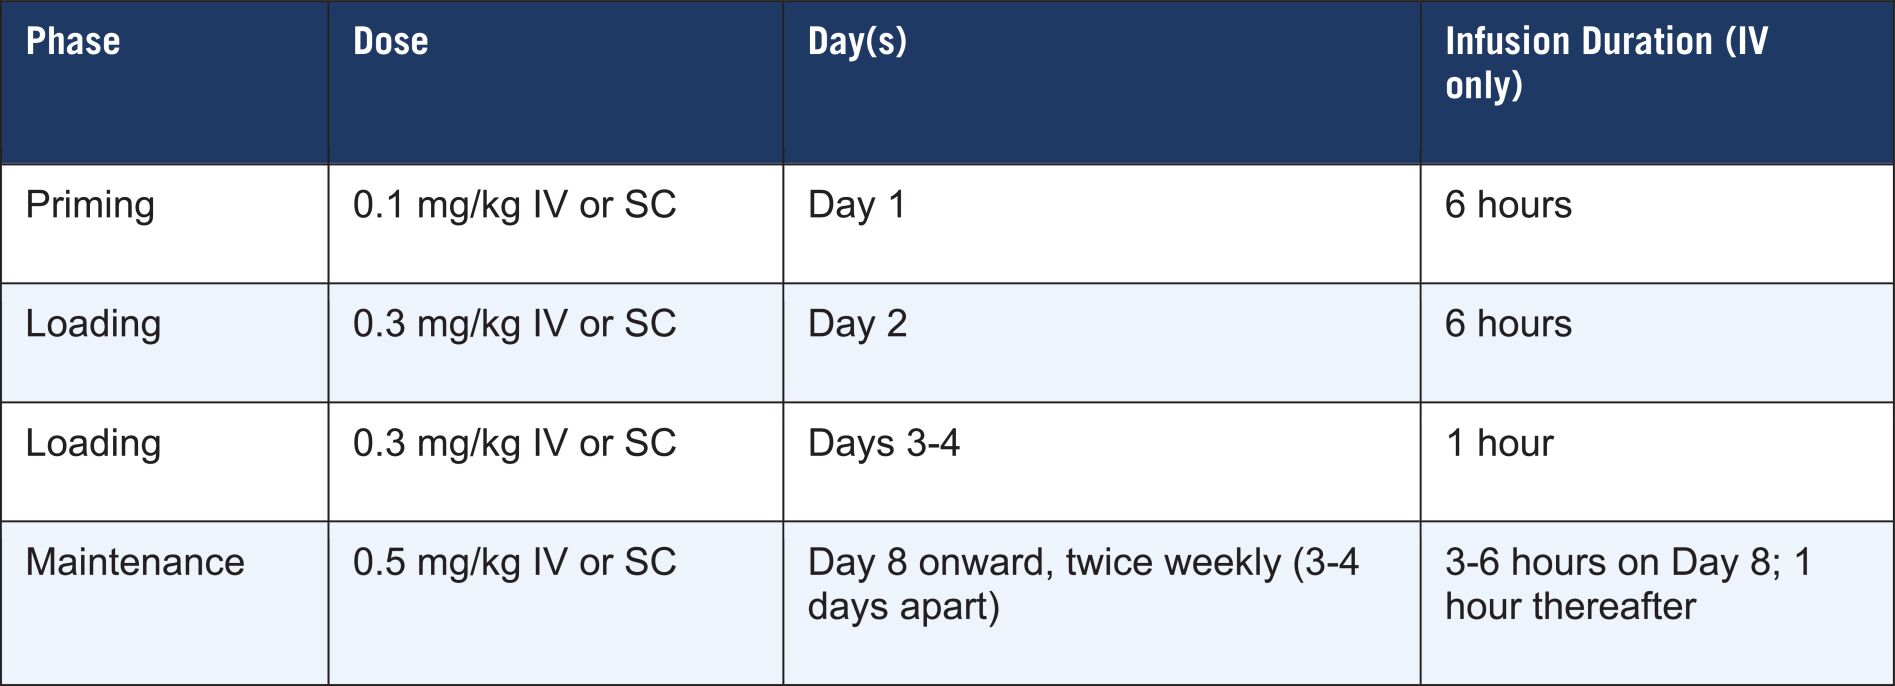
125
Table of Contents
Participants initially receive dexamethasone as premedication and background therapy, with subsequent tapering as clinically appropriate. SOC chemotherapy for the underlying malignancy in mHLH patients is permitted and encouraged. Patients aged 12 years and older who continue to derive clinical benefit at the conclusion of the 12-week treatment course may enter an extension treatment phase at the discretion of their physician and with the approval of the Medical Monitor. Long-term survival follow-up occurs every 12 weeks for up to two years following the end of treatment.
SURPASS is being conducted at approximately 30 clinical sites across the United States and Europe, with enrollment initiated in the second half of 2025. COMPASS is also being conducted across the United States and Europe. We expect to complete enrollment in our registrational program in the second half of 2027. Subject to positive results, we intend to submit for regulatory approval of ipsoprubart.
Significant Unmet Need and Commercial Opportunity in sHLH
We believe ipsoprubart has the potential to meaningfully improve outcomes for patients with sHLH and transform the current treatment paradigm. In our Phase 1b trial, treatment with ipsoprubart in frontline sHLH patients was associated with rapid clinical responses, hospital discharge, and improved survival. We believe these outcomes have the potential to provide meaningful clinical and health economic benefits by reducing the morbidity and mortality associated with uncontrolled hyperinflammation and enabling patients to transition more rapidly from inpatient to outpatient care.
Substantial and Growing Addressable Patient Population
Based on our analysis of U.S. claims data using HLH-associated ICD codes, we estimate that approximately 12,600 patients were diagnosed with sHLH in the United States in 2025. Diagnosed incidence increased at an estimated compound annual growth rate of 11.5% from 2021 through 2025. We believe this estimate understates the true burden of disease because it does not capture patients who may be treated without assignment of an HLH-associated ICD code or misdiagnosed or undiagnosed patients.
Recent studies suggest that the prevalence of HLH-associated hyperinflammation may be substantially greater than currently recognized. For example, one study reported that approximately 24% of patients with diffuse large B-cell lymphoma (DLBCL) met OHI diagnostic criteria. DLBCL is the most common aggressive lymphoma, and we estimate it accounts for approximately 20,000 new U.S. cases annually. If applied across the broader population of patients with hematologic malignancies, this observation could imply a substantially larger addressable patient population. Additional studies have suggested that approximately 4% to 5% of patients presumed to have sepsis may in fact have underlying sHLH, further highlighting the potential for underdiagnosis.
We believe diagnosed incidence will continue to increase as awareness of sHLH grows, underlying triggering conditions become more prevalent, particularly with the expanding use of immune-effector cell therapies, and simplified diagnostic frameworks, including the OHI index, facilitate earlier recognition and diagnosis.
Readily Identifiable Patients and Highly Concentrated Treater Base
Patients with sHLH typically present with persistent fever and other manifestations of rapidly progressive systemic inflammation. The severity and acuity of symptoms generally lead patients to seek urgent medical care, resulting in immediate intervention, laboratory evaluation, and involvement of hematologist-oncologists, who are typically responsible for the diagnosis and management of sHLH both in the outpatient and inpatient settings.
The physician base responsible for treating sHLH is highly concentrated. Hematologist-oncologists serve as the primary treating physicians for the majority of patients. Based on third-party claims analyses, we estimate that approximately 64% of sHLH patients in the United States are treated at academic medical centers and that 200 healthcare organizations account for approximately 51% of all diagnosed sHLH patients. We believe this concentration may support an efficient commercial launch of ipsoprubart through a hematology-oncology focused field organization targeting a relatively small number of physicians and treating institutions.
High Unmet Need with No Broadly Approved Therapy
There is currently no established SOC and no approved therapy for the treatment of sHLH broadly. Physicians rely on a heterogeneous combination of corticosteroids, chemotherapy, cytokine inhibitors, and other off-label therapies, yet
126
Table of Contents
mortality outcomes have remained poor despite two decades of advances in supportive care and related therapeutic areas. We believe an effective frontline therapy capable of rapidly controlling hyperinflammation and improving survival could address a significant unmet need across the broad sHLH patient population.
Potentially Favorable Orphan Disease Pricing and Access Dynamics
We believe ipsoprubart, if approved, may offer a compelling value proposition for patients, physicians, and payers. Clinically meaningful early response and hospital discharge rates have the potential to reduce progression to multiorgan failure, prolonged intensive care utilization, and other costly complications of uncontrolled disease. In addition, we anticipate that the majority of the treatment course would occur in the outpatient setting following stabilization of the acute disease onset. Given the severity of sHLH, the absence of approved therapies that broadly address the disease, and the potential clinical and health economic benefits of treatment, we believe ipsoprubart may benefit from favorable orphan disease pricing and market access dynamics, if approved.
Ipsoprubart: Expansion into Hematologic Malignancies
In an exploratory analysis of eight patients treated with ipsoprubart across our Phase 1b trial and Expanded Access Program, we observed a 100% objective tumor response rate and an 88% CR rate by Week 14. One patient treated with ipsoprubart as a monotherapy experienced a CR with respect to both sHLH and the underlying heavily refractory T cell lymphoma. The patient was alive as of the last follow-up, more than one year post treatment. We believe T/NK cell malignancies remain areas of significant unmet medical need, particularly in the relapsed/refractory setting where available therapies demonstrate low CR rates, limited durability, and poor long-term outcomes. We believe these diseases represent a substantial commercial opportunity, and we estimate a diagnosed incidence rate of more than 13,000 annually in the United States across the T/NK cell malignancy subtypes. Based on the anti-tumor activity observed in the patients from the Phase 1b trial and Expanded Access Program, we have initiated a Phase 1 clinical trial of ipsoprubart in relapsed/refractory T/NK cell malignancies and expect initial data in the second half of 2027.
Ipsoprubart: Clinical Data in Patients with Hematologic Malignancies
Treatment with ipsoprubart in our Phase 1b trial and Expanded Access Program demonstrated anti-tumor activity in sHLH patients with T cell and B cell lymphomas whose tumor responses were evaluated in an exploratory analysis, including patients whose cancer was highly refractory to multiple prior lines of therapy.
We believe ipsoprubart may induce anti-tumor activity through two complementary mechanisms: (1) direct depletion of SIRPg-expressing malignant T cells in patients with T cell malignancies, in which the malignant cells themselves may express the target antigen, particularly tumors with an activated T cell phenotype, and (2) depletion of SIRPa/ß1-expressing myeloid cells and SIRPg-expressing T cells in the tumor microenvironment, potentially reducing immunosuppressive signaling and supporting an enhanced anti-tumor immune response.
A total of eight patients with sHLH and underlying T cell and B cell lymphomas were evaluable for objective tumor response during the early course of ipsoprubart treatment—seven enrolled in our Phase 1b trial and one through an Expanded Access Program. The seven patients treated in the Phase 1b trial, six of whom had relapsed/refractory lymphoma, received ipsoprubart in combination with concomitant lymphoma-directed therapies, including gemcitabine and oxaliplatin, romidepsin, multi-agent chemotherapy, and loncastuximab tesirine. The patient treated under the Expanded Access Program received ipsoprubart monotherapy.
127
Table of Contents
The lymphoma subtypes of these eight patients and their best response on treatment with ipsoprubart are summarized in the table below. Of these eight patients, 100% achieved an objective tumor response by Week 14, as assessed by the investigator using standard criteria appropriate for the underlying lymphoma subtype, during their ipsoprubart treatment course. Notably, 88% (7 of 8) of patients achieved a CR by Week 14.
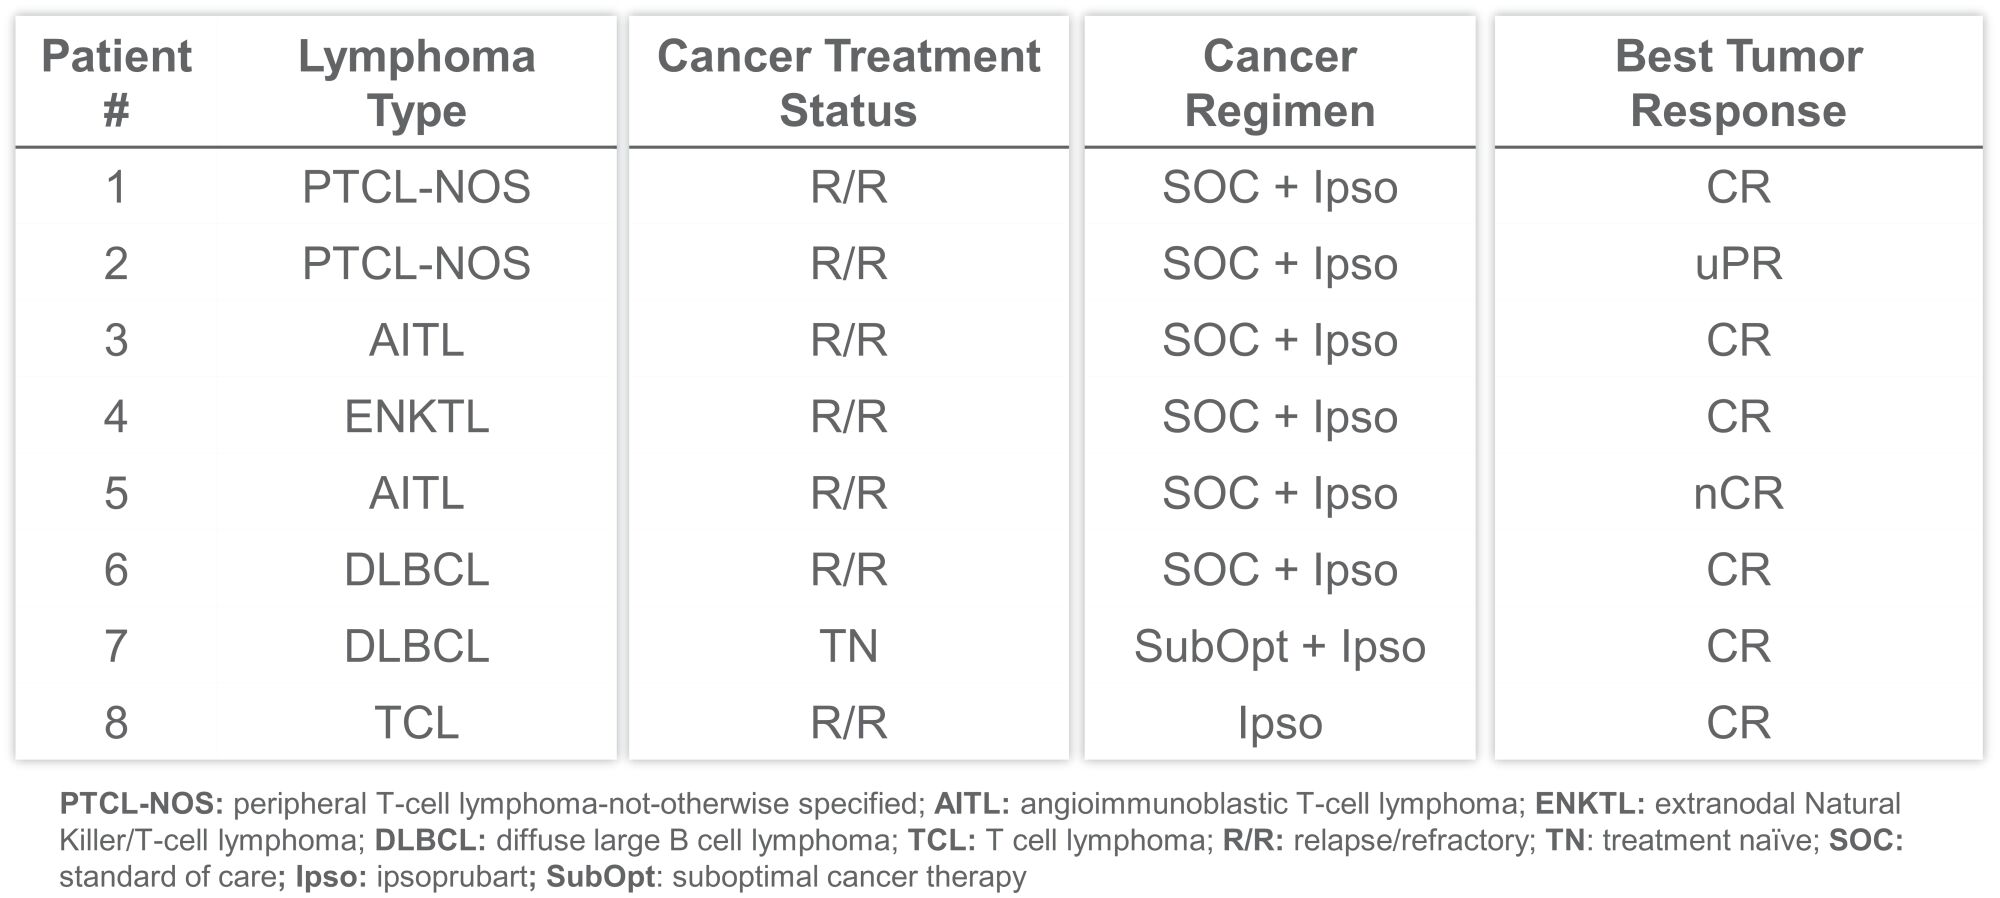
Responsecategories were assigned at the investigator’s discretion as there was no pre-specified analysis in the protocol. In general, the assigned response definitions have the following meaning: CR (Complete Response): No evidence of disease. nCR (Nodal Complete Response): No evidence of residual disease in lymph nodes (typically based on PET or PET/CT imaging), without necessarily requiring confirmation of bone marrow clearance. PR (Partial Response): A partial reduction in disease burden, typically meeting the criteria for a ≥50% reduction in measurable disease. Unsustained PR: A patient who initially achieves a partial response but subsequently loses that response (e.g., due to disease progression) before achieving a complete response or maintaining the PR.
Case Study: Durable Complete Response in Patient with sHLH Triggered by DLBCL
The patient was a 45-year-old male with sHLH triggered by treatment-naïve T cell/histiocyte-rich DLBCL treated in the Phase 1b trial. He received a 12-week course of ipsoprubart alongside less than one month of cyclophosphamide and rituximab treatment, as well as dexamethasone background therapy. Of note, the patient elected for a substantially less intensive chemotherapy regimen and did not elect for standard-of-care chemotherapy regimen for DLBCL (for example, rituximab in combination with cyclophosphamide, doxorubicin, vincristine, and prednisone or R-CHOP).
The patient achieved an mCR for sHLH and a CR for their DLBCL at Week 8, as demonstrated by the PET-CT images below where the left scan shows the patient’s DLBCL disease activity at baseline and the right scan shows normalization of the PET-CT scan after 8 weeks of ipsoprubart treatment.
128
Table of Contents
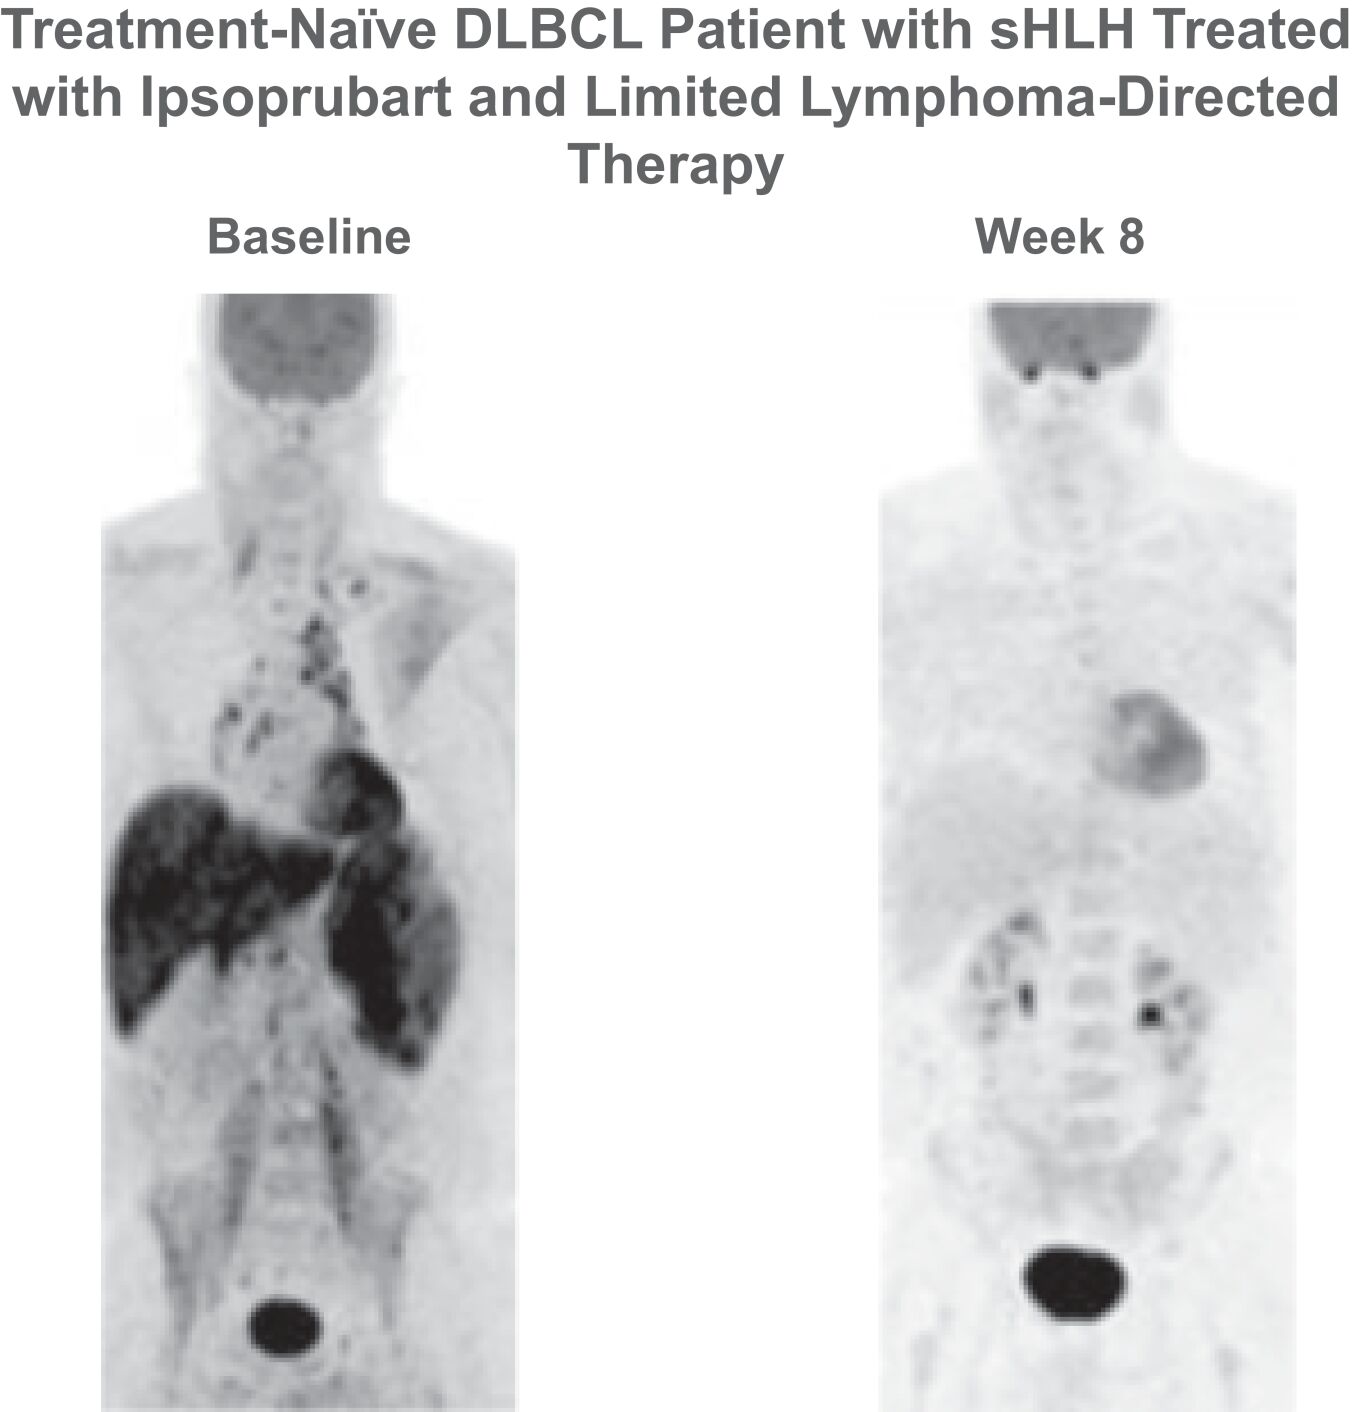
The patient’s DLBCL CR was durable for more than one year as of the patient’s last follow-up. We believe this case is informative because the patient received less than one month of limited-intensity lymphoma therapy, with no anthracycline exposure. In the context of DLBCL, durable CRs are typically associated with multi-agent combination chemotherapy. Taken together, these factors suggest that ipsoprubart may have contributed meaningfully to the durable CR observed in this patient.
Case Study: Durable Complete Response in a Patient with T Cell Lymphoma After Ipsoprubart Monotherapy
This patient is the same individual described previously in the subsection titled “—Complete Response in mHLH Patient Treated with Frontline Ipsoprubart Monotherapy.” This 26-year-old male patient presented with sHLH triggered by refractory cutaneous cytotoxic CD8+ T cell lymphoma and had received more than five lines of prior anti-cancer therapy before accessing ipsoprubart through an Expanded Access Program. Additional details regarding the tumor response are provided below.
The patient experienced a complete tumor response following ipsoprubart monotherapy. As shown in the left PET-CT image below, the patient had a massively enlarged spleen of 28.5 cm extending to the pelvis at baseline. Following completion of ipsoprubart treatment, the spleen length decreased to approximately 13.7 cm, as shown in the right PET-CT image taken at Week 13, representing an estimated spleen volume reduction of over 65%.
129
Table of Contents
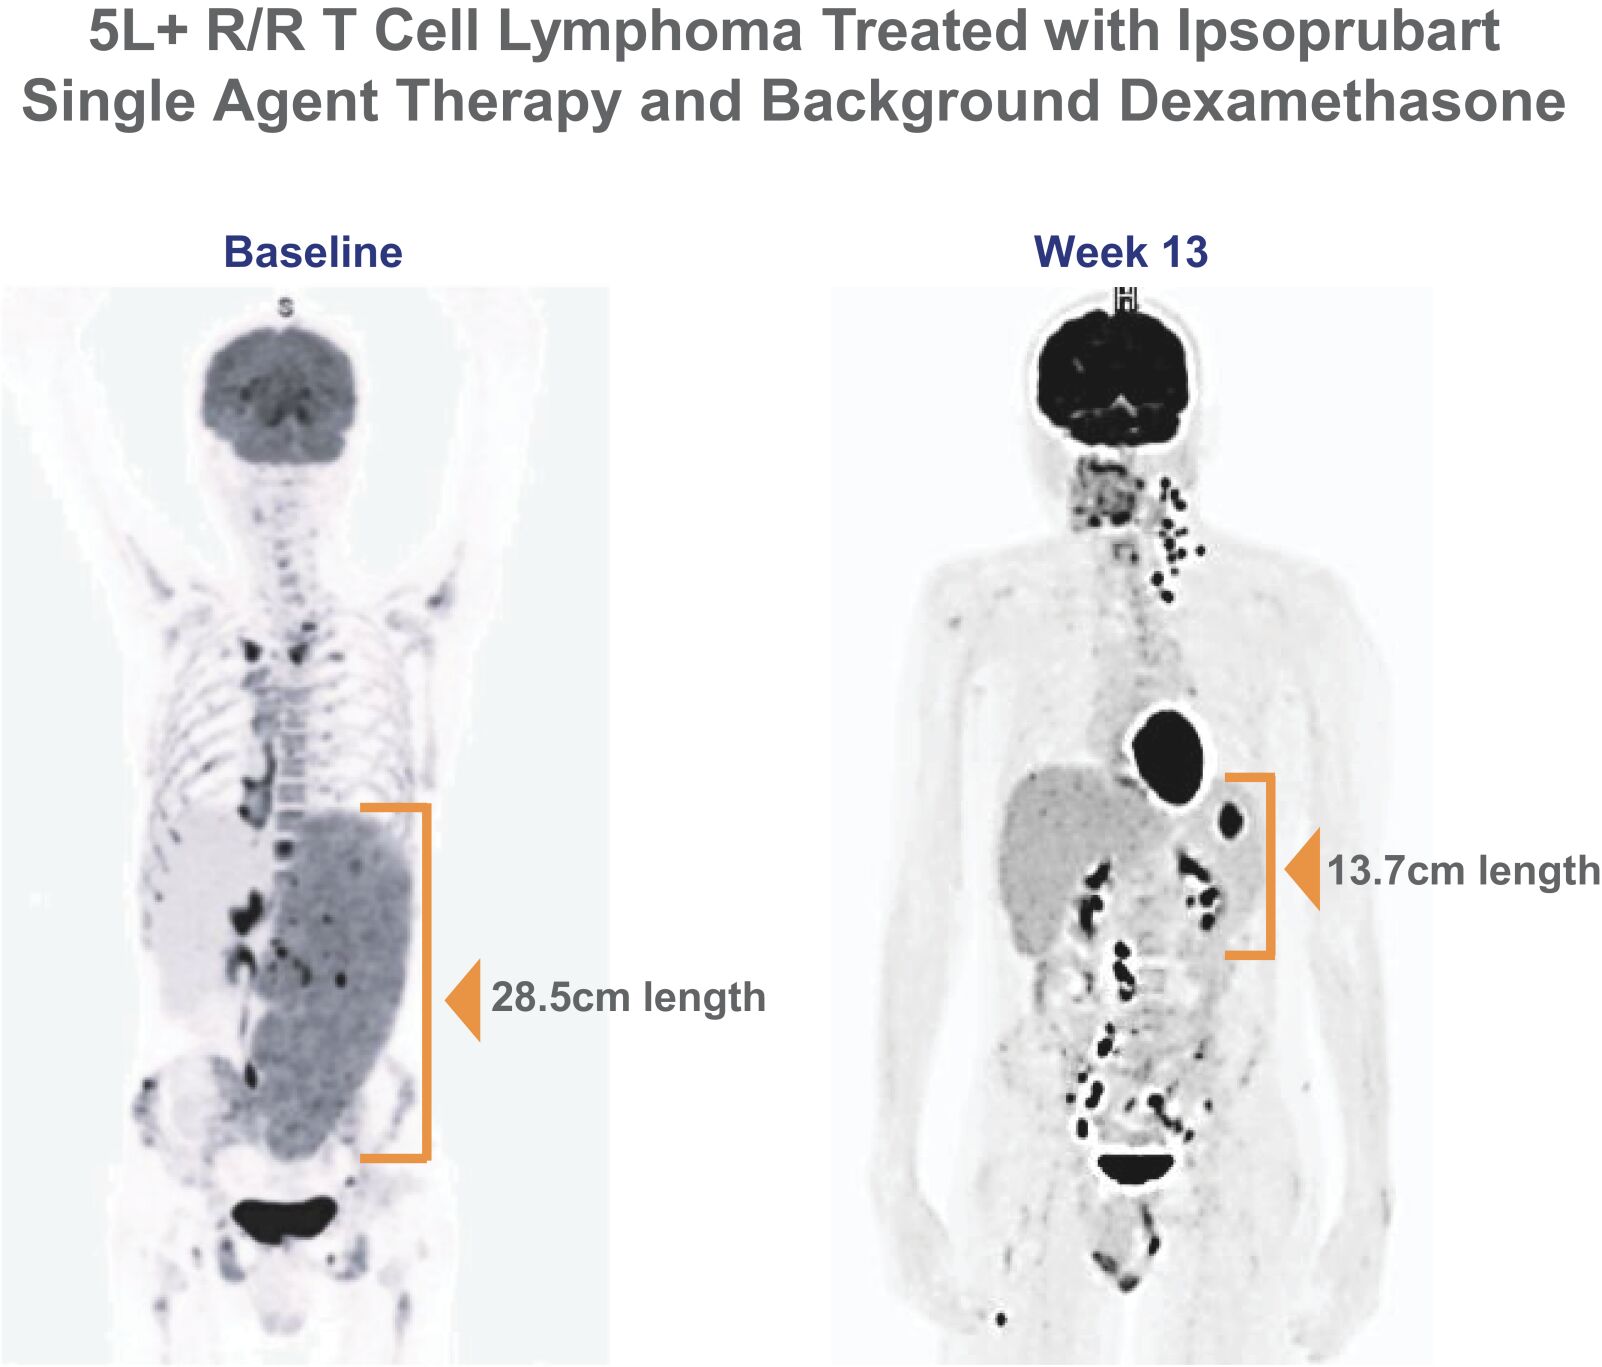
The response was further evidenced by a marked improvement in symptoms and laboratory findings. The patient’s blood cell counts recovered, bone marrow was cleared of lymphoma, and lymph nodes showed reduced fluorodeoxyglucose uptake on PET-CT imaging. The patient remained alive more than one year following ipsoprubart treatment as of the most recent follow-up.
We believe these patient cases support the potential of ipsoprubart to deliver meaningful single-agent activity as well as synergistic effects in combination with a broad range of cancer-directed therapies by targeting both tumor cells and the tumor microenvironment. The depth and durability of the responses observed support further investigation of ipsoprubart in patients with hematologic malignancies.
Our Ongoing Phase 1 Clinical Trial of Ipsoprubart in T/NK Cell Malignancies
Based on the anti-tumor activity observed in our Phase 1b trial and Expanded Access Program, we submitted an IND to the FDA, which was allowed to proceed in 2025. We have initiated a Phase 1, two-part, multicenter trial to evaluate ipsoprubart in participants at least 18 years old with relapsed/refractory T/NK cell malignancies following any line of prior therapy who are eligible for investigational treatments. The trial will evaluate the safety, tolerability, PK, PD, and preliminary activity of ipsoprubart. Key efficacy endpoints include overall response, disease control, and overall survival. The trial will begin with an initial dose finding component (Part 1, Phase 1a), enrolling approximately 6 to 18 participants (up to maximum of 24 participants) to identify up to two dosing regimens with an acceptable safety profile.
130
Table of Contents
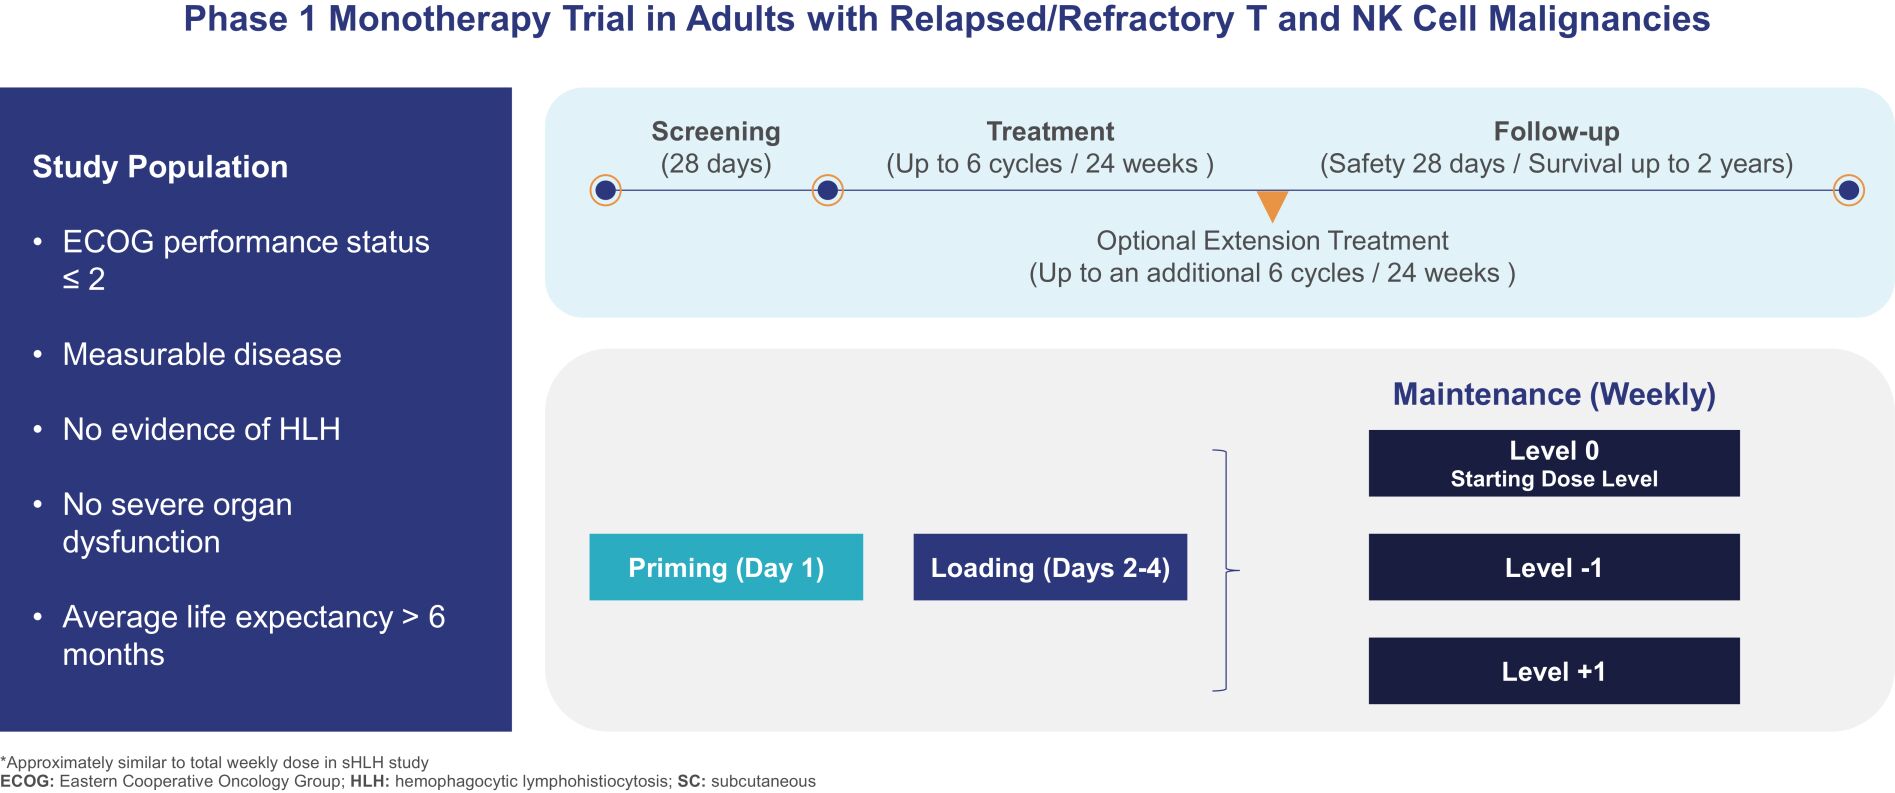
Upon completion of Part 1, the trial will proceed to the cohort expansion phase (Part 2, Phase 1b) to further evaluate these regimens. We expect initial data from this trial in the second half of 2027.
Significant Commercial Opportunity in T/NK Cell Malignancies with Strategic Synergies in sHLH
T/NK cell malignancies remain areas of significant unmet medical need, particularly in the relapsed/refractory setting, where currently available therapies are often associated with low complete response rates, limited durability of response, and poor long-term outcomes. Historically reported CR rates to standard chemotherapy in relapsed/refractory peripheral T cell lymphoma can range from approximately 10% to 15%. We believe these diseases represent a substantial commercial opportunity, and we estimate a diagnosed incidence rate of more than 13,000 annually in the United States across the T/NK cell malignancy subtypes. In addition, the physicians who diagnose and treat these malignancies are largely the same hematologist-oncologists who manage patients with sHLH, creating potential commercial and operational synergies between applicable indications for ipsoprubart. We believe ipsoprubart’s mechanism of targeting activated myeloid cells and T cells, together with the anti-tumor activity observed to date, supports continued evaluation in these malignancies.
ELA822: SIRPg-Specific Antibody for Chronic Immune and Inflammatory Diseases
ELA822, our second product candidate, is a fully human IgG1k monoclonal antibody from our SIRP-targeted antibody portfolio, which was selected for its SIRPg-specificity, with no binding to SIRPa or SIRPß1, and contains a canonical IgG1 Fc intended for repeated, long-term administration. By selectively engaging cells expressing high levels of SIRPg, ELA822 is designed to deplete activated, pathogenic T cells while sparing naïve and regulatory myeloid cells and T cells. We believe this target profile is well suited for a broad range of chronic T cell-mediated immune and inflammatory diseases where durable efficacy and long-term tolerability are important therapeutic objectives. Nonclinical data for ELA822 demonstrated selective depletion of activated T cells in vitro and activity in humanized models of T cell-driven diseases in vivo, and ELA822 was well tolerated in NHPs. We have obtained regulatory clearance and initiated a Phase 1 clinical trial of ELA822 in healthy volunteers in Europe, with data expected in the first half of 2027, followed by initiation of a Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which is expected in the middle of 2027, subject to regulatory approval.
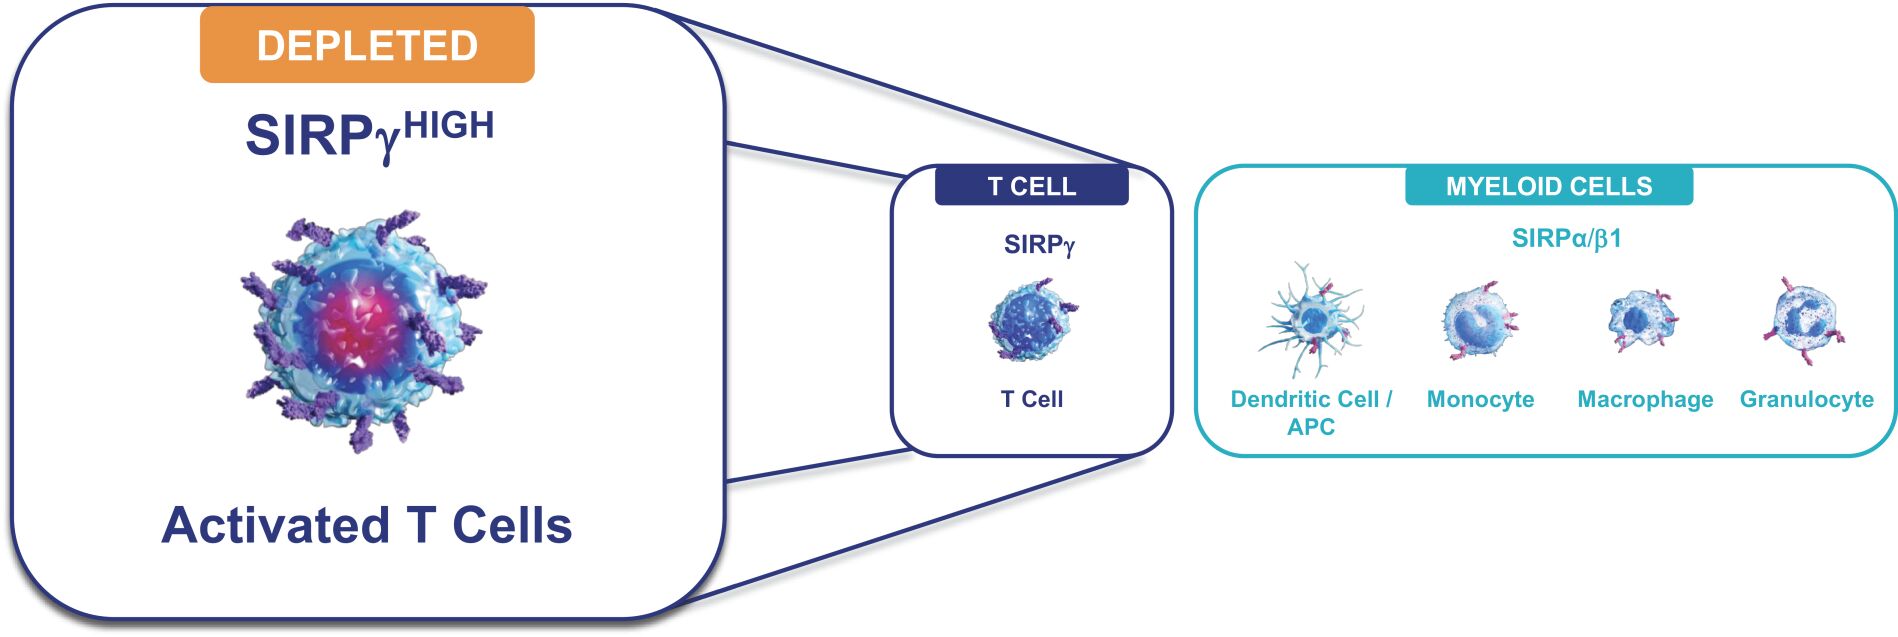
131
Table of Contents
Nonclinical Studies
The pharmacodynamic effect of ELA822 administration was evaluated in NHPs. As in humans, SIRPg expression in NHPs is largely restricted to T cells and it is not typically detected on myeloid cells. However, SIRPg expression has been shown to be approximately 2.6-fold higher in NHPs, as measured by median fluorescence intensity (MFI), and only a small proportion of peripheral blood T cells are reported to be SIRPg-negative cells (approximately 11%). Thus, although NHPs represent a pharmacologically relevant species for evaluating ELA822, PD effects may be more pronounced than in humans because of the higher level and broader distribution of SIRPg expression. Weekly administration of ELA822 of in NHPs resulted in a dose dependent decrease in lymphocyte counts, as measured by a complete blood count (CBC) with differential. The maximal reduction of approximately 50% was consistent with the proportion of T lymphocytes present in the peripheral blood of NHPs. Immunophenotyping further demonstrated a dose-dependent decrease in CD3+ T cells, with a nadir of approximately 75%. Analysis of SIRPg expression on the residual T cell population was consistent with preferential depletion of T cells expressing the highest levels of SIRPg.
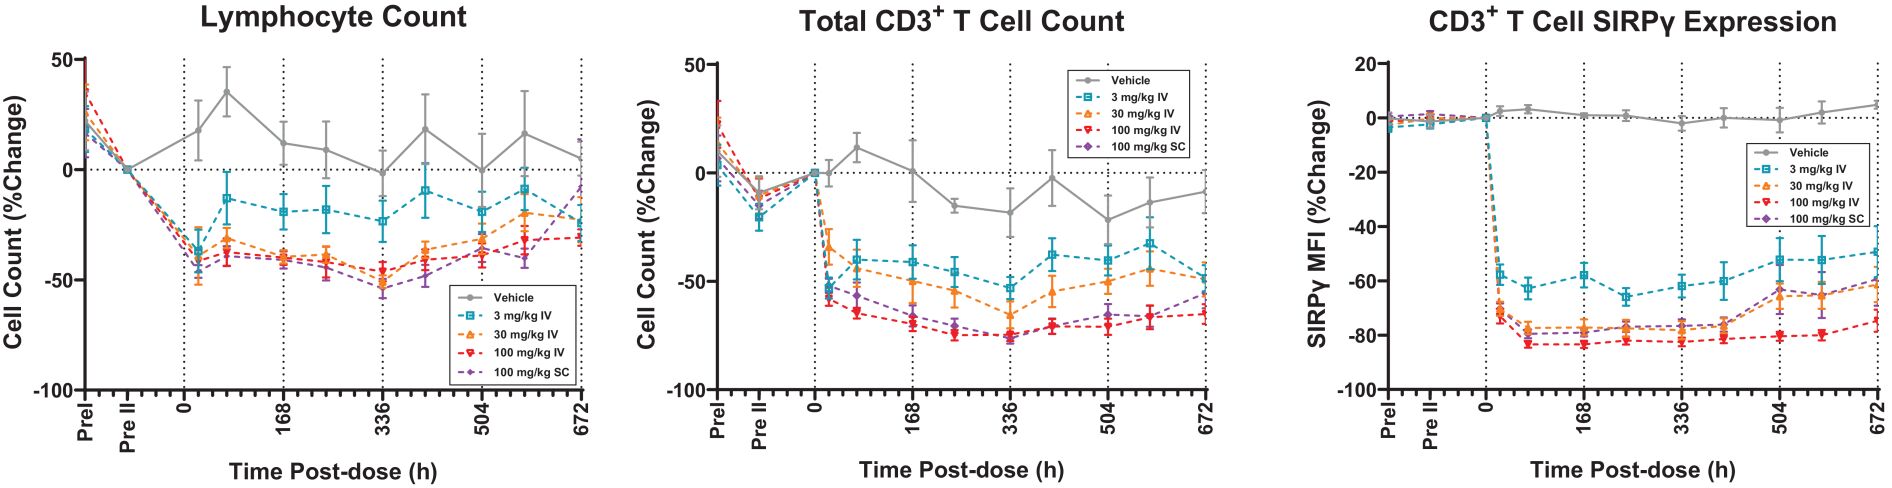
We have evaluated the activity of ELA822 in a humanized mouse model of giant cell arteritis (GCA), a serious inflammatory condition that causes swelling in the lining of arteries, in collaboration with the Mayo Clinic. ELA822 demonstrated a statistically significant reduction (p<0.05) in CD4+ T cell infiltration of the temporal artery wall relative to an isotype control antibody at the highest dose. ELA822 also produced statistically significant reductions (p<0.05) in lesional expression of the T cell effector cytokines IFNg and IL-21 compared to the isotype control at the highest dose. Taken together, we believe these findings demonstrate that SIRPg plays a pivotal role in GCA lesion formation and establish in vivo proof-of-concept for the ability of ELA822 to deplete pathogenic T cell populations and be a potential disease modifying treatment for GCA.
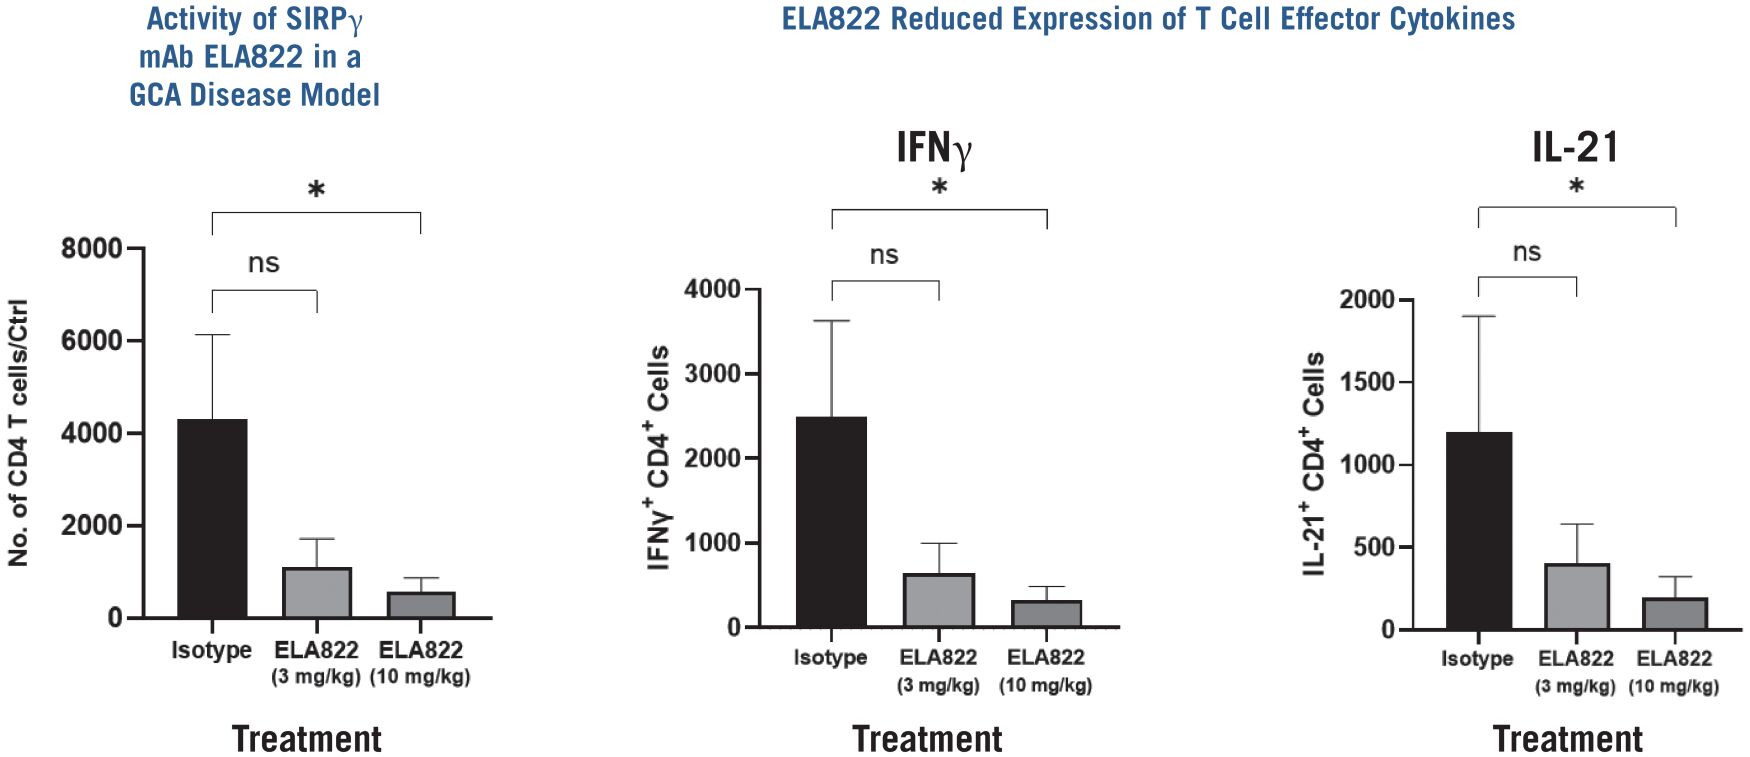
GCA: Giant Cell Arteritis; IFNg: Interferon-gamma
In a humanized mouse model of graft-versus-host disease (GvHD), administration of a SIRPg targeted monoclonal antibody was associated with depletion of activated HLA-DR+/CD38+ T cells and produced clinically meaningful benefits relative to treatment with an isotype control antibody. ELA822-treated animals demonstrated a reduction in mean GvHD score at Day 28, from seven to four on a 1 to 10 scale where a total daily score of 0 was considered healthy, while a score of 10 indicated severe GvHD. ELA822 treatment also prevented the loss in body weight associated with GvHD and resulted in improved survival of animals through Day 30 of follow-up.
132
Table of Contents
We have also completed a 28-day IND-enabling toxicology study of ELA822 in NHPs. Repeat IV dosing was well tolerated at doses up to 100 mg/kg with no drug-related adverse clinical or pathologic observations, establishing a no-observed-adverse-effect level of 100 mg/kg IV. ELA822 demonstrated SC bioavailability of greater than 90% in NHPs, which, if observed in humans, may support convenient outpatient SC administration for chronic immune and inflammatory diseases. Additionally, the PD effects of ELA822 observed in NHPs were highly selective for the T cell compartment.
Phase 1 Clinical Program to Assess ELA822
We have initiated a Phase 1 clinical trial to evaluate the safety, tolerability, PD, and PK of ELA822 in healthy volunteers. This trial is a randomized, placebo-controlled, single-ascending-dose study, with sentinel dosing at each dose level. We plan to select the starting dose for evaluation in T cell-mediated diseases based on emerging safety, PK, and PD data from this trial. Based on regulatory feedback, we expect to finalize our trial design for our planned Phase 1/2 clinical trial in patients with T cell-mediated immune disorders, which we expect to initiate in the middle of 2027, subject to regulatory approval.
Potential Breadth of Application in T Cell-Mediated Diseases
We believe ELA822 may have application across a broad range of T cell-mediated immune and inflammatory diseases in which pathogenic T cell subsets express SIRPg at levels that may support selective depletion. The figure below illustratively depicts the constellation of diseases in which activated or pathogenic T cells have been implicated.
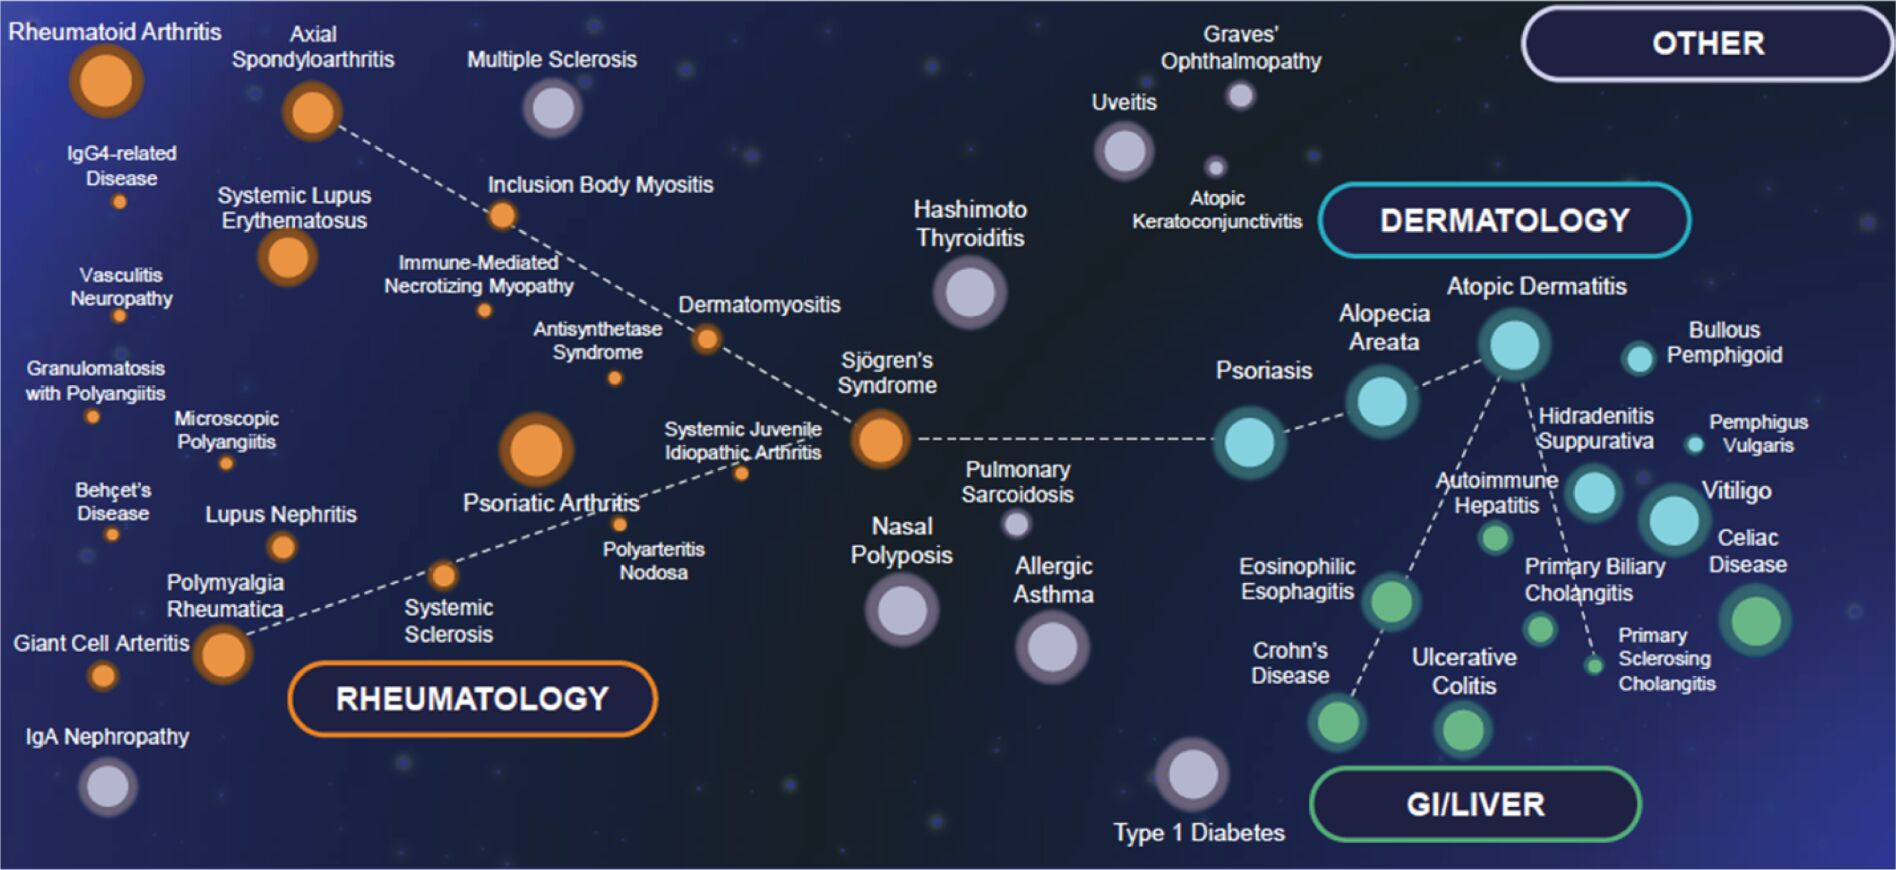
Manufacturing
We oversee and manage third party contract manufacturing organizations (CMOs) to support development and manufacture of protein biologic product candidates for our clinical trials. We expect our strategy to use CMOs will enable us to maintain a more efficient infrastructure, avoiding the necessity to acquire our own manufacturing facility and equipment, while simultaneously enabling us to focus our expertise on clinical development and the potential future commercialization of our products. Currently, we rely on and have agreements with multiple third-party CMOs to manufacture and supply drug substance (DS) and drug product (DP) for our clinical trials. In the event we advance any of our drug candidates through Phase 3 clinical trials, we anticipate the need to enter into a manufacture and supply agreement with, and transfer DS and DP manufacture to, one or more larger third-party CMOs with whom we would also likely enter into commercial supply agreements with prior to any potential regulatory approval if any of our drug candidates are commercialized. The DS and DP for our drug candidates are manufactured via conventional biopharmaceutical processing procedures, employing commonly used and commercially available excipients and packaging materials. The DS and DP are required to be manufactured in compliance with current good manufacturing practices (cGMP) and comparable foreign requirements. The process, equipment and methods employed for manufacture and analysis are consistent with standard biologics synthesis or pharmaceutical production, and are transferable to a range of manufacturing facilities, if needed.
133
Table of Contents
Competition
The biopharmaceutical industry is characterized by rapidly evolving science and technology, intense competition, and a strong emphasis on intellectual property. We face, and will continue to face, substantial competition from biotechnology and pharmaceutical companies of all sizes, public and private research institutions, and governmental agencies. Many of our competitors have substantially greater financial, technical, and personnel resources than we do, more experience in the discovery, development, manufacturing, and commercialization of pharmaceutical products, and more established commercial infrastructure. These competitors also compete with us in recruiting and retaining qualified scientific and management personnel, establishing clinical trial sites, enrolling patients in clinical trials, and acquiring or in-licensing technologies that are complementary to, or potentially necessary for, our programs. Mergers and acquisitions in the biopharmaceutical industry may result in even more resources being concentrated among a smaller number of competitors. Smaller or earlier-stage companies may also prove to be significant competitors, particularly through collaborative arrangements with large, established companies. Any of our competitors may develop or commercialize therapies that are more effective, safer, more convenient, or less expensive than our product candidates, may obtain regulatory approval before we do, or may develop intellectual property that limits our ability to develop or commercialize our product candidates.
Ipsoprubart in sHLH
There is currently no established SOC and no therapies approved for the treatment of sHLH broadly. Patients with sHLH are commonly treated with high dose corticosteroids, chemotherapy, cytokine inhibitors, and other off-label therapies that non-selectively suppress the immune response or target individual cytokines. Emapalumab (Gamifant, marketed by Swedish Orphan Biovitrum AB) is approved by the FDA for primary HLH and MAS triggered by Still’s disease but is not approved for sHLH broadly. We will also compete with any investigational therapies in development now or in the future for sHLH and related hyperinflammatory conditions.
Ipsoprubart in Hematologic Malignancies
For our planned development of ipsoprubart in T/NK cell malignancies, we expect to compete with currently approved therapies for T cell malignancies. These therapies include alemtuzumab (Campath, marketed by Sanofi Genzyme), belinostat (Beleodaq, marketed by Acrotech Biopharma LLC), bexarotene (Targretin, marketed by Bausch Health Companies Inc.), brentuximab vedotin (Adcetris, marketed by Pfizer Inc. and Takeda Pharmaceutical Company Limited), denileukin diftitox (Lymphir, marketed by Citius Pharmaceuticals, Inc.), mogamulizumab (Poteligeo, marketed by Kyowa Kirin Co., Ltd.), nelarabine (Arranon, marketed by Novartis Pharmaceuticals Corporation), pralatrexate (Folotyn, marketed by Acrotech Biopharma LLC), romidepsin (Istodax, marketed by Bristol-Myers Squibb Company), valemetostat (Ezharmia, marketed by Daiichi Sankyo, Inc.), and vorinostat (Zolinza, marketed by Merck Sharp & Dohme LLC). We will also compete with investigational therapies in development now or in the future for T/NK cell malignancies.
ELA822 in Chronic Immune and Inflammatory Diseases
ELA822 is being developed for chronic T cell-mediated immune and inflammatory diseases, an area in which there is well-established competition from multiple classes of approved therapies such as JAK inhibitors, TYK2 inhibitors, IL-4 and IL-13 antagonists, IL-17 inhibitors, and IL-23 inhibitors. We will also compete with investigational therapies in development now or in the future for chronic T cell-mediated immune and inflammatory diseases, such as programs targeting OX40, CD94, and KLRG1.
Intellectual Property
Our commercial success depends in large part on our ability to obtain and maintain patent protection and other proprietary and intellectual property protection in the U.S. and other countries for our current and future product candidates. We seek to protect our proprietary position by, among other methods, filing U.S. and foreign patents and patent applications. We will also seek to rely on regulatory protection afforded through inclusion in expedited development and review, data exclusivity, market exclusivity, and patent term extensions where available.
Generally, issued patents have a term of 20 years from the earliest claimed non-provisional filing date in the applicable country, subject to the payment of applicable maintenance, renewal, annuity, and other governmental fees. In certain instances in the United States, patent terms can be adjusted to recapture a portion of delay by the USPTO in examining the patent application, or extended to account for patent term effectively lost as a result of the FDA regulatory review process, subject to applicable statutory and regulatory requirements and a cap of 14 years of patent protection remaining from the date of product approval by the FDA, or both. Similar provisions are available in Europe and certain other jurisdictions to extend the term of a patent covering an approved drug.
134
Table of Contents
As of August 26, 2026, our patent estate, which consists of Company-owned patents and applications, includes 2 issued U.S. patents, 5 pending U.S. non-provisional patent applications, 1 issued foreign patent, 70 pending foreign patent applications, and 5 pending international patent applications filed under the Patent Cooperation Treaty (PCT patent application), not yet nationalized. The foreign pending applications are in various countries or regions outside of the U.S., including Australia, Brazil, Canada, China, Hong Kong, India, Israel, Japan, Mexico, New Zealand, Singapore, South Africa, South Korea, Taiwan, and countries within the European Patent Convention and the Eurasian Patent Organization. These patents and applications include filings directed toward our SIRPa/ß1/g targeted antibodies, including ipsoprubart, and SIRPg specific antibodies, including ELA822. The issued patents, and any patents that may issue from the pending patent applications are expected to expire between 2041-2046, without taking into account any potential patent term adjustments, extensions or terminal disclaimers, and assuming payment of all appropriate maintenance, renewal, annuity, and other governmental fees. We expect to file additional patent applications in support of current and future product candidates, as well as their uses.
SIRPa/ß1/g Antibodies
We own patents and applications directed to novel SIRPa/ß1/g targeted antibodies (pan-SIRP antibodies), and methods of making, and using these antibodies. As of August 26, 2026, our pan-SIRP antibodies patent portfolio includes 2 issued U.S. patents, 3 pending U.S. non-provisional patent applications, 1 issued foreign patent, 39 pending foreign patent applications, and 2 pending PCT patent applications, which have not yet been nationalized. The issued U.S. patents provide coverage for antibodies that bind SIRPa, SIRPß1, and SIRPg, including ipsoprubart, and are expected to expire in 2041, without taking into account any potential patent term adjustments, extensions or terminal disclaimers, and assuming payment of all appropriate maintenance, renewal, annuity and other governmental fees. Other patents that may issue from these pending applications are expected to expire between 2041-2046, without taking into account any potential patent term adjustments, extensions or terminal disclaimers, and assuming payment of all appropriate maintenance, renewal, annuity, and other governmental fees.
SIRPg Antibodies
In addition to the pan-SIRP antibodies patents and applications outlined above, we also own patent applications directed to SIRPg-specific antibodies, including ELA822, providing compositions of matter, and methods of making, and methods of use. As of August 26, 2026, these filings include 1 pending U.S. non-provisional patent application, 18 pending foreign patent applications, and 2 pending PCT patent applications, which have not yet been nationalized. Any patents that issue from these pending patent applications are expected to expire between 2042-2045, without taking into account any potential patent term adjustments, extensions, or terminal disclaimers, and assuming payment of all appropriate maintenance, renewal, annuity, and other governmental fees.
Although we will continue to file patent applications and to seek to maximize the scope of our patent protection for all of our programs, the coverage claimed in a patent application can be significantly reduced before a patent is issued, and its scope can be reinterpreted and even challenged after issuance, therefore, we cannot provide any assurance that any patents will issue from our pending or future applications or that any issued patents will adequately protect our current or future product candidates. Without patent protection for our current or future product candidates, we may be open to competition from third parties. For more information, see the section titled “Risks Related to Our Intellectual Property.”
We maintain and are seeking registered trademarks. As of August 26, 2026, we own 1 registered trademark on the company name and 1 pending trademark application in the United States. These numbers may grow as our portfolio continues to evolve and expand.
We also protect the trade secrets, know-how, and other proprietary information underlying our product candidates. To help protect our trade secrets, know-how, and other proprietary information, we rely, in part, upon non-disclosure, confidentiality, and non-competition agreements with our commercial partners, collaborators, employees, and consultants, and invention assignment agreements with our employees. We limit the disclosure of our trade secrets to employees and third-party service providers who have a need to know such information in order to perform their responsibilities. We also seek to preserve the integrity and confidentiality of our data and trade secrets by maintaining the physical security of our premises and physical and electronic security of our information technology systems. While we aim to ensure we have agreements in place to help protect our trade secrets and confidential know-how, these agreements may be breached, and we may not have adequate remedies for any resulting breach. In addition, our trade secrets may otherwise become known or be independently discovered by competitors. To the extent that our commercial partners, collaborators, employees, and consultants use intellectual property owned by others in their work for us, disputes may arise as to the rights in related or resulting patents, patent applications, know-how, inventions, and other intellectual property. For more information, see the section titled “Risks Related to Our Intellectual Property.”
135
Table of Contents
Obtaining patents does not guarantee our right to practice or commercialize the patented technology. Any patent we own or may own in the future may be challenged, circumvented or invalidated by third parties. In addition, third parties may have or in the future obtain rights to patents that could be used to prevent or attempt to prevent us from commercializing our current or future product candidates. If third parties prepare and file patent applications in the United States or other jurisdictions that also claim technology to which we have rights, we may have to participate in proceedings in the USPTO, such as interference or derivation proceedings, in the federal courts, or in similar proceedings in other jurisdictions to determine the priority of invention, or potential validity, enforceability, or infringement. For more information see the section titled “Risks Related to Our Intellectual Property.”
Government Regulation
Government authorities in the United States, at the federal, state, and local level, and in other countries and jurisdictions, extensively regulate, among other things, the research, development, testing, manufacture, quality control, approval, packaging, storage, recordkeeping, labeling, advertising, promotion, distribution, marketing, post-approval monitoring and reporting, and import and export of pharmaceutical products. The processes for obtaining regulatory approvals in the United States and in foreign countries and jurisdictions, along with subsequent compliance with applicable statutes and regulations and other regulatory authorities, require the expenditure of substantial time and financial resources.
FDA Approval Process
In the United States, biological products, such as those we are developing, are subject to extensive regulation by the FDA. The Federal Food, Drug, and Cosmetic Act (FDC Act), the Public Health Service Act (PHS Act), and other federal and state statutes and regulations govern, among other things, the research, development, testing, manufacture, storage, recordkeeping, approval, labeling, promotion and marketing, distribution, post-approval monitoring and reporting, sampling, and import and export of pharmaceutical products. Biological products used for the prevention, treatment, or cure of a disease or condition of a human being are subject to regulation under the FDC Act, except the section of the FDC Act which governs the approval of New Drug Applications (NDAs). Biological products are approved for marketing under provisions of the PHS Act, via a Biologics License Application (BLA). However, the application process and requirements for approval of BLAs are very similar to those for NDAs. Failure to comply with applicable U.S. requirements may subject a company to a variety of administrative or judicial sanctions, such as a clinical hold, FDA refusal to approve a pending BLA, warning or untitled letters, product recalls, product seizures, total or partial suspension of production or distribution, injunctions, fines, civil penalties, and criminal prosecution.
The process required by the FDA before a drug or biologic may be marketed in the United States generally involves the following:
| ∎ | completion of certain nonclinical laboratory tests, animal studies and formulation studies in accordance with Good Laboratory Practice (GLPs) regulations and other applicable requirements and; |
| ∎ | submission to the FDA of an Investigational New Drug Application (IND), which must become effective before human clinical trials may begin; |
| ∎ | approval by an independent institutional review board (IRB), or ethics committee at each clinical site before each trial may be initiated; |
| ∎ | performance of adequate and well-controlled human clinical trials in accordance with Good Clinical Practice regulations (GCPs) to evaluate the safety, purity and potency, of the product candidate for its intended use; |
| ∎ | preparation and submission to the FDA of a BLA; |
| ∎ | satisfactory completion of an FDA advisory committee review, if applicable; |
| ∎ | satisfactory completion of an FDA inspection of the manufacturing facility or facilities at which the drug is produced to assess compliance with current Good Manufacturing Practice requirements (cGMPs) to assure that the facilities, methods and controls are adequate to preserve the biological product’s continued safety, purity, and potency; |
| ∎ | satisfactory completion of potential inspection of selected clinical investigation sites to assess compliance with GCPs; and |
| ∎ | FDA review and approval of the BLA to permit commercial marketing of the product for particular indications for use in the United States. |
Nonclinical Studies
Nonclinical studies include laboratory evaluation of product chemistry, formulation, and toxicity, as well as animal trials to assess the characteristics and potential safety and efficacy of the product. The conduct of the nonclinical
136
Table of Contents
tests must comply with federal regulations and requirements, including GLP requirements for certain animal studies. The results of nonclinical testing are submitted to the FDA as part of an IND along with other information, including information about product chemistry, manufacturing, and controls (CMC) and a proposed clinical trial protocol. Long-term nonclinical tests, such as animal tests of reproductive toxicity and carcinogenicity, may continue after the IND is submitted. An IND automatically becomes effective 30 days after receipt by the FDA, unless before that time the FDA raises concerns or questions related to one or more proposed clinical trials and places the trial on a full or partial clinical hold. In such a case, the IND sponsor and the FDA must resolve any outstanding concerns before the clinical trial can begin or begin as proposed. Submission of an IND therefore may or may not result in FDA allowance to begin a clinical trial.
Clinical Trials
Clinical trials involve the administration of the investigational biologic to participants including healthy volunteers or patients under the supervision of a qualified investigator. Clinical trials must be conducted: (i) in compliance with any applicable federal regulations; (ii) in compliance with good clinical practices (GCPs), which are regulations and standards meant to protect the rights and health of patients and to define the roles of clinical trial sponsors, administrators, and monitors; as well as (iii) under protocols detailing the objectives of the trial, the parameters to be used in monitoring safety, and any safety effectiveness criteria to be evaluated. Each protocol involving testing on U.S. patients and subsequent protocol amendments must be submitted to the FDA as part of the IND. While the IND is active, progress reports summarizing the results, if known, of the clinical trials and nonclinical studies performed since the last progress report, among other information, must be submitted at least annually to the FDA, and written IND safety reports must be submitted to the FDA and investigators in certain circumstances, including for serious and unexpected suspected adverse events, findings from other studies suggesting a significant risk to humans exposed to the same or similar drugs, findings from animal or in vitro testing suggesting a significant risk to humans, and any clinically important increased incidence of a serious suspected adverse reaction compared to that listed in the protocol or investigator brochure.
The FDA may order the temporary, or permanent, discontinuation of a clinical trial at any time, or impose other sanctions, if it believes that the clinical trial either is not being conducted in accordance with FDA requirements or presents an unacceptable risk to the clinical trial patients; in such a case a sponsor may voluntarily suspend or discontinue a study. The study protocol and informed consent information for patients in clinical trials must also be submitted to IRBs or ethics committees overseeing clinical sites, for approval. An IRB may also require the clinical trial at the site to be halted, either temporarily or permanently, for a variety of reasons, including failure to comply with the IRB’s requirements or if there is a finding that patients are exposed to an unacceptable health risk, or may impose other conditions. Some studies also include oversight by an independent group of qualified experts organized by the clinical trial sponsor, which may be known as a data safety monitoring board.
A sponsor who wishes to conduct a clinical trial outside of the United States may, but need not, obtain FDA authorization to conduct the clinical trial under an IND. If a foreign clinical trial is not conducted under an IND, the sponsor may submit data from the clinical trial to the FDA in support of an IND or a BLA. The FDA will accept the results from a well-designed and well-conducted foreign clinical trial not conducted under an IND if the clinical trial was conducted in accordance with GCPs and the FDA is able to validate the data through an onsite inspection if deemed necessary.
Clinical trials to support BLAs for regulatory approval are typically conducted in three sequential phases, but the phases may be combined or overlap.
| ∎ | In Phase 1, the initial introduction of the biological product candidate into healthy volunteers or patients, the product candidate is tested to assess safety, dosage tolerance, metabolism, pharmacokinetics, pharmacological actions, side effects associated with drug exposure, and, if possible, early evidence on effectiveness. |
| ∎ | Phase 2 usually involves trials in a limited patient population with the target disease or condition to preliminarily evaluate the effectiveness of the biologic product candidate for a particular indication, determine optimal dose and regimen for further development, and to identify common adverse effects and safety risks. |
| ∎ | Phase 3 trials are subsequently undertaken in an expanded patient population to obtain additional information about clinical efficacy and safety in a larger number of patients, typically at geographically dispersed clinical trial sites, to permit the FDA to evaluate the overall benefit-risk relationship of the biologic product candidate and to provide adequate information for the labeling of the product. |
137
Table of Contents
Post-approval trials, sometimes referred to as Phase 4 studies, may be conducted after initial marketing approval. These trials are used to gain additional experience from the treatment of patients in the intended therapeutic indication. In certain instances, the FDA may mandate the performance of Phase 4 clinical trials as a condition of approval of a BLA.
The sponsor of an investigational product in a Phase 2 or Phase 3 clinical trial for a serious or life-threatening disease is required to make available, such as by posting on its website, its policy on evaluating and responding to requests for expanded access to such investigational drug.
Concurrent with clinical trials, companies usually complete additional animal studies and also must develop additional information about the chemistry and physical characteristics of the drug or biologic as well as finalize a process for manufacturing the product in commercial quantities in accordance with cGMP requirements. The manufacturing process must be capable of consistently producing quality batches of the product and, among other things, companies must develop methods for testing the identity, strength, quality, potency, and purity of the final product. Additionally, appropriate packaging must be selected and tested, and stability studies must be conducted to demonstrate that the investigational medicines do not undergo unacceptable deterioration over their shelf life.
FDA Review Process
Assuming successful completion of all required testing in accordance with all applicable regulatory requirements, the product candidate sponsor prepares and submits a BLA to the FDA seeking approval to market the biologic for one or more indications. FDA approval of the BLA is required before marketing and distribution of the product may begin in the United States. The BLA must include the results of all preclinical, clinical, and other testing and a compilation of data relating to the product’s pharmacology and CMC. The cost of preparing and submitting a BLA is substantial. The submission of most BLAs is additionally subject to a substantial application user fee. Under an approved BLA, the applicant is also subject to an annual program fee. These fees typically increase annually. A BLA for a biologic that has been designated as an orphan drug is not subject to an application fee, unless the BLA includes an indication for other than a rare disease or condition.
The FDA has 60 days from its receipt of a BLA to conduct a preliminary review and determine whether the application will be filed based on the FDA’s threshold determination that the BLA is sufficiently complete to permit substantive review. If the FDA determines the application is incomplete because it does not on its face contain required information, the FDA may refuse to file the application and request additional information rather than file the BLA. In this event, the BLA must be resubmitted with the additional information. The resubmitted application also is subject to preliminary review before the FDA files it. Once the application is filed, the FDA begins an in-depth review. The FDA reviews a BLA to determine, among other things, whether the product candidate is safe, pure and potent and the facility in which it is manufactured, processed, packed or held meets standards designed to assure the product’s continued safety, purity and potency.
The FDA has agreed to certain performance goals in the review of BLAs. Standard-review applications have a goal of being reviewed within ten months of the date the FDA files the BLA; applications classified as Priority Review have a goal of being reviewed within six months of the date the FDA files the BLA. A BLA can be classified for Priority Review when the FDA determines the biologic product candidate has the potential to treat a serious or life-threatening condition and, if approved, would be a significant improvement in safety or effectiveness compared to available therapies. The FDA does not always meet its PDUFA goal dates for standard and priority BLAs, and the review process for both standard and priority reviews may be extended by the FDA for three additional months to consider certain late-submitted information or information deemed a “major amendment” to the BLA submission.
The FDA may also refer applications for novel biologic products, as well as biologic products that present difficult questions of safety or efficacy, to an advisory committee—typically a panel that includes clinicians and other experts—for review, evaluation, and a recommendation as to whether the BLA should be approved. The FDA is not bound by the recommendation of an advisory committee, but it often follows such recommendations.
Before approving a BLA, the FDA will typically inspect one or more clinical sites to assure compliance with GCPs. Additionally, the FDA will generally inspect the facility or the facilities at which the biologic product is manufactured. The FDA will not approve the product unless compliance with cGMP is satisfactory and adequate to assure consistent production of the product within required specifications.
138
Table of Contents
After the FDA evaluates the BLA and completes any clinical and manufacturing site inspections, it issues either an approval letter or a complete response letter. A complete response letter generally outlines the deficiencies in the BLA and may require substantial additional clinical or nonclinical testing, or other information, in order for the FDA to reconsider the BLA for approval. If, or when, those deficiencies have been addressed to the FDA’s satisfaction in a resubmission of the BLA, the FDA will issue an approval letter. The FDA has committed to reviewing such resubmissions in two or six months depending on the type of information included. Even if such data and information are submitted, the FDA may decide that the BLA does not satisfy the criteria for approval.
An approval letter authorizes commercial marketing and distribution of the biologic with specific prescribing information for specific indications. As a condition of BLA approval, the FDA may require a risk evaluation and mitigation strategy (REMS) to help ensure that the benefits of the biologic outweigh the potential risks. REMS can include medication guides, communication plans for healthcare professionals, and elements to assure safe use (ETASU). ETASU can include, but are not limited to, special training or certification for prescribing or dispensing, dispensing only under certain circumstances, special monitoring, and the use of patient registries. The requirement for a REMS can materially affect the potential market and profitability of the product. Moreover, the FDA may require substantial post-approval testing, sometimes referred to as Phase 4 testing, and surveillance to monitor the product’s safety or efficacy.
Once granted, product approvals may be withdrawn if compliance with regulatory standards is not maintained, or problems are identified following initial marketing. Changes to some of the conditions established in an approved BLA, including changes in indications, labeling, or manufacturing processes or facilities, require submission and FDA approval of a new BLA or BLA supplement before the change can be implemented. A BLA supplement for a new indication typically requires clinical data similar to that in the original application, and the FDA uses the same procedures and actions in reviewing BLA supplements as it does in reviewing BLAs.
Disclosure of Clinical Trial Information
Sponsors of clinical trials of FDA-regulated products, including biologics, are required to register and disclose certain clinical trial information on ClinicalTrials.gov. Information related to the product, patient population, phase of investigation, study sites and investigators, and other aspects of the clinical trial is then made public as part of the registration. Sponsors are also obligated to disclose the results of their clinical trials after completion. Disclosure of the results of these trials can be delayed in certain circumstances for up to two years after the date of completion of the trial. Competitors may use this publicly available information to gain knowledge regarding the progress of development programs.
Additional Controls for Biologics
To help reduce the increased risk of the introduction of adventitious agents, the PHS Act emphasizes the importance of manufacturing controls for products whose attributes cannot be precisely defined. The PHS Act also provides authority to the FDA to immediately suspend biologics licenses in situations where there exists a danger to public health, to prepare or procure products in the event of shortages and critical public health needs, and to authorize the creation and enforcement of regulations to prevent the introduction or spread of communicable diseases within the United States.
After a BLA is approved, the product may also be subject to official lot release as a condition of approval. As part of the manufacturing process, the manufacturer is required to perform certain tests on each lot of the product before it is released for distribution. If the product is subject to official release by the FDA, the manufacturer submits samples of each lot of product to the FDA together with a release protocol showing a summary of the lot manufacturing history and the results of all of the manufacturer’s tests performed on the lot. The FDA may also perform certain confirmatory tests on lots of some products, such as viral vaccines, before allowing the manufacturer to release the lots for distribution. In addition, the FDA conducts laboratory research related to the regulatory standards on the safety, purity, potency, and effectiveness of biological products. After approval of a BLA, biologics manufacturers must address any safety issues that arise, are subject to recalls or a halt in manufacturing, and are subject to periodic inspection after approval.
Orphan Drug Designation
Under the Orphan Drug Act, the FDA may grant orphan designation to a drug or biologic intended to treat patients with a rare disease or condition, which is a disease or condition that affects fewer than 200,000 individuals in the
139
Table of Contents
United States, or more than 200,000 individuals in the United States for which there is no reasonable expectation that the cost of developing and making available in the United States a drug or biologic for this type of disease or condition will be recovered from sales in the United States for that drug or biologic. Orphan drug designation must be requested before submitting a BLA. After the FDA grants Orphan Drug Designation, the generic identity of the therapeutic agent and its potential orphan use are disclosed publicly by the FDA. The Orphan Drug Designation does not convey any advantage in, or shorten the duration of, the regulatory review or approval process.
If a product that has Orphan Drug Designation subsequently receives the first FDA approval for the rare disease or condition for which it has such designation, the product is entitled to orphan drug exclusive approval (or exclusivity), which means that the FDA may not approve any other applications, including a full BLA, to market the same drug for the same approved indication or use for seven years, except in limited circumstances, such as a showing of clinical superiority to the product with orphan drug exclusivity within the relevant indication or use, or if the holder of the orphan drug exclusivity cannot assure the availability of sufficient quantities of the orphan drug to meet the needs relating to the approved indication or use of patients with the relevant disease or condition. Orphan drug exclusivity does not prevent the FDA from approving a different drug or biologic for the same indication or use, or the same drug or biologic for a different indication or use. Among the other benefits of Orphan Drug Designation are tax credits for certain research and a waiver of the BLA application user fee.
Expedited Development and Review Programs
The FDA offers a number of programs intended to expedite the development or review of a marketing application for an investigational biologic. For example, the Fast Track designation program is intended to expedite or facilitate the process for developing and reviewing product candidates that meet certain criteria. Specifically, investigational biologics are eligible for Fast Track designation if they are intended to treat a serious or life-threatening disease or condition and demonstrate the potential to address unmet medical needs for the disease or condition. The sponsor of a Fast Track product candidate has opportunities for more frequent interactions with the applicable FDA review team during product development and, once a BLA is submitted, the application may be eligible for priority review. With regard to a Fast Track product candidate, the FDA may consider for review sections of the BLA on a rolling basis before the complete application is submitted, if the sponsor provides a schedule for the submission of the sections of the BLA, the FDA agrees to accept sections of the NDA or BLA and determines that the schedule is acceptable, and the sponsor pays any required user fees upon submission of the first section of the BLA.
A product candidate intended to treat a serious or life-threatening disease or condition may also be eligible for Breakthrough Therapy designation to expedite its development and review. A product candidate can receive Breakthrough Therapy designation if preliminary clinical evidence indicates that the product candidate, alone or in combination with one or more other drugs or biologics, may demonstrate substantial improvement over existing therapies on one or more clinically significant endpoints, such as substantial treatment effects observed early in clinical development. The designation includes all of the Fast Track program features, as well as more intensive FDA interaction and guidance beginning as early as Phase 1 and an organizational commitment to expedite the development and review of the product candidate, including involvement of senior managers.
In addition, a BLA may be eligible for priority review if the product candidate is designed to treat a serious condition, and if approved, would provide a significant improvement in safety or efficacy compared to available therapies for such disease or condition. The FDA will attempt to direct additional resources to the evaluation of a BLA designated for priority review in an effort to facilitate the review. Priority review designation means the FDA’s goal is to take action on the marketing application within six months of the 60-day filing date, as compared to ten months for standard review under current PDUFA goals.
In addition, depending on the design of the applicable clinical trials, a product candidate may be eligible for accelerated approval. Specifically, biologics intended to treat serious or life-threatening diseases or conditions may be eligible for accelerated approval upon a determination that the product candidate has an effect on a surrogate endpoint that is reasonably likely to predict clinical benefit, or on a clinical endpoint that can be measured earlier than irreversible morbidity or mortality, that is reasonably likely to predict an effect on irreversible morbidity or mortality or other clinical benefit, taking into account the severity, rarity, or prevalence of the condition and the availability or lack of alternative treatments. As a condition of approval, the FDA generally requires that a sponsor of a biologic receiving accelerated approval perform adequate and well-controlled confirmatory clinical trials and may
140
Table of Contents
require that such confirmatory trials be underway prior to granting accelerated approval. Biologics receiving accelerated approval may be subject to expedited withdrawal procedures if the sponsor fails to conduct the required confirmatory trials in a timely manner or if such trials fail to verify the predicted clinical benefit. In addition, the FDA requires pre-approval of promotional materials as a condition of accelerated approval, which could adversely impact the timing of the commercial launch of the product.
Fast Track designation, Breakthrough Therapy designation, priority review, and accelerated approval do not change the standards for approval, but they may expedite the development or approval process. Even if a product candidate qualifies for one or more of these programs, the FDA may later decide that the product no longer meets the conditions for qualification or decide that the time period for FDA review or approval will not be shortened.
U.S. Patent Term Restoration, Marketing Exclusivity, and Biosimilars
Depending upon the timing, duration, and specifics of FDA approval of our product candidates, some of our U.S. patents may be eligible for limited patent term extension under the Drug Price Competition and Patent Term Restoration Act of 1984 (the Hatch-Waxman Amendments). The Hatch-Waxman Amendments provide for a patent term extension of up to five years as compensation for patent term lost during the FDA regulatory review process, subject to applicable statutory and regulatory requirements. Patent term extension, however, cannot extend the remaining term of a patent beyond a total of 14 years from the product’s approval date. The patent term extension period is generally one half the time between the effective date of an IND and the submission date of a BLA, plus the entire period between the submission date of a BLA and the approval of that application, subject to a maximum extension of five years and the 14-year post-approval cap. If the patent selected for extension was issued after the start of the review period, only the portion of the review period occurring after patent issuance is considered in calculating the extension. The review period will be reduced by any time during which the applicant failed to exercise due diligence. Only one patent applicable to an approved drug is eligible for such an extension and such patent may be extended only once, only those claims covering the approved drug, a method for using it, or a method for manufacturing it may be extended, and the application for the extension must be submitted prior to the expiration of the patent. Such application must be submitted within 60 days of the product’s approval. The USPTO, in consultation with the FDA, determines the eligibility of a patent for patent term extension and the length of such extension, if any, and there can be no assurance that any of our U.S. patents will be eligible for, or receive, the patent term extension, or that any such extension will be granted for the full period sought.
The Biologics Price Competition and Innovation Act of 2009 (BPCIA) created an abbreviated approval pathway for biological products shown to be highly similar to or interchangeable with an FDA-licensed reference biological product. Biosimilarity sufficient to reference a prior FDA-approved product requires that there be no differences in conditions of use, route of administration, dosage form, and strength, and no clinically meaningful differences between the biological product and the reference product in terms of safety, purity, and potency. Biosimilarity may be shown through analytical studies, toxicity studies, and a clinical trial or trials, unless the Secretary of Health and Human Services waives a required element. A biosimilar product may be deemed interchangeable with a previously approved product if it meets the higher hurdle of demonstrating that it can be expected to produce the same clinical results as the reference product and, for products administered multiple times, the biologic and the reference biologic may be switched after one has been previously administered without increasing safety risks or risks of diminished efficacy relative to exclusive use of the reference biologic. Under most state laws, interchangeable products may be used in place of the reference biological product.
A reference biologic is granted 12 years of marketing exclusivity from the time of first licensure, or BLA approval, of the reference product, and no application for a biosimilar or interchangeable product referencing the reference product can be accepted by the FDA for four years from the date of first licensure of the reference product. In addition, the approval of a biosimilar product may not be made effective by the FDA until 12 years from the date on which the reference product was first licensed. During this 12-year period of exclusivity, another company may still develop and receive approval of a competing biologic, so long as their BLA does not rely on the reference product or sponsor’s data or submit the application as a biosimilar application. “First licensure” typically means the initial date the particular product at issue was licensed in the United States. Date of first licensure does not include the date of licensure of (and a new period of exclusivity is not available for) a biological product if the licensure is for a supplement for the biological product or for a subsequent application by the same sponsor or manufacturer of the biological product (or licensor, predecessor in interest, or other related entity) for a change (not including a
141
Table of Contents
modification to the structure of the biological product) that results in a new indication, route of administration, dosing schedule, dosage form, delivery system, delivery device or strength, or for a modification to the structure of the biological product that does not result in a change in safety, purity or potency. The first biologic product submitted under the biosimilar abbreviated approval pathway that is licensed as interchangeable with the reference product has exclusivity against the approval of other interchangeable biologics for the same condition of use for the lesser of (i) one year after first commercial marketing of the first interchangeable biosimilar, (ii) 18 months after the first interchangeable biosimilar is approved if there is no patent challenge under the BPCIA patent-litigation provisions, (iii) 18 months after resolution of a lawsuit under the BPCIA patent-litigation provisions over the patents of the reference biologic in favor of the first interchangeable biosimilar applicant, or (iv) 42 months after the first interchangeable biosimilar’s application has been approved if a patent lawsuit under the BPCIA patent-litigation provisions is ongoing within the 42-month period.
Pediatric Information
Under the Pediatric Research Equity Act (PREA), BLAs or supplements to BLAs must contain data to assess the safety and effectiveness of the biological product candidate for the claimed indications in all relevant pediatric subpopulations and to support dosing and administration for each pediatric subpopulation for which the biological product is deemed safe, pure and potent. The FDA may grant full or partial waivers or deferrals for submission of data. A deferral may be granted for several reasons, including a finding that the drug is ready for approval for use in adults before pediatric clinical trials are complete or that additional safety or effectiveness data needs to be collected before the pediatric clinical trials begin. Unless otherwise required by regulation, PREA does not apply to any biological product for a disease or condition for which orphan designation has been granted, though still applies to any non-orphan-designated indication.
The Best Pharmaceuticals for Children Act (BPCA) provides a six-month extension of non-patent exclusivity and certain patent terms for a biologic if certain conditions are met. Conditions for exclusivity include the FDA’s determination that information relating to the use of a new biologic in the pediatric population may produce health benefits in that population, the FDA making a written request for pediatric studies, and the applicant agreeing to perform, and reporting on, the requested studies within the statutory time frame. Applications under the BPCA are treated as priority applications, with all of the benefits that designation confers; however, a sponsor need not obtain approval for the use of the biologic within the relevant pediatric disease or condition to obtain pediatric exclusivity.
Post-Approval Requirements
Once a BLA is approved, a product will be subject to certain post-approval requirements. For instance, the FDA closely regulates the post-approval marketing and promotion of biologics, including standards and regulations for direct-to-consumer advertising, off-label promotion, industry-sponsored scientific and educational activities and promotional activities involving the internet. Biologics may be marketed only for the approved indications and in accordance with the provisions of the approved labeling.
The FDA strictly regulates marketing, labeling, advertising, and promotion of biologics that are placed on the market. Advertising and promotion of biologics must be in compliance with the FDC Act and its implementing regulations and only for the approved indications and in a manner consistent with the approved labeling. The FDA and other agencies actively enforce the laws and regulations prohibiting the promotion of off-label uses, and a company that is found to have improperly promoted off-label uses may be subject to significant liability, including investigation by federal and state authorities. Physicians may prescribe legally available products for uses that are not described in the product’s labeling and that differ from those tested by us and approved by the FDA. Such off-label uses are common across medical specialties. Physicians may believe that such off-label uses are the best treatment for many patients in varied circumstances. The FDA does not regulate the behavior of physicians in their choice of treatments. The FDA does, however, restrict manufacturer’s communications on the subject of off-label use of their products.
Adverse event reporting and submission of periodic reports is required following FDA approval of a BLA. The FDA also may require post-marketing testing, including Phase 4 testing, REMS, and surveillance to monitor the effects of an approved product, or the FDA may place conditions on an approval that could restrict the distribution or use of the product. In addition, quality control, biological product manufacture, packaging, and labeling procedures must continue to conform to cGMP after approval. Biologic manufacturers and certain of their subcontractors are required to register their establishments with the FDA and certain state agencies. Registration with the FDA subjects entities
142
Table of Contents
to periodic unannounced inspections by the FDA, during which the agency inspects a biologic product’s manufacturing facilities to assess compliance with cGMP. Accordingly, manufacturers must continue to expend time, money, and effort in the areas of production and quality-control to maintain compliance with cGMP. Regulatory authorities may withdraw product approvals or request product recalls if a company fails to comply with regulatory standards, if it encounters problems following initial marketing, or if previously unrecognized problems are subsequently discovered.
Other potential consequences include, among other things:
| ∎ | Form 483s, restrictions on the manufacturing of the product, product recalls or withdrawal of the product from the market; |
| ∎ | warning or untitled letters or holds on clinical trials; |
| ∎ | refusal of the FDA to accept new marketing applications or approve pending applications or supplements to approved applications, or suspension or revocation of product approvals or licensures; |
| ∎ | product seizure or detention, or refusal to permit the import or export of products; |
| ∎ | consent decrees, corporate integrity agreements, debarment, or exclusion from federal healthcare programs; |
| ∎ | mandated modification of promotional materials and labeling and the issuance of corrective information; |
| ∎ | the issuance of safety alerts, Dear Healthcare Provider letters, press releases and other communications containing warnings or other safety information about the product; or |
| ∎ | injunctions or the imposition of fines or civil or criminal penalties. |
Healthcare Laws and Regulations
Sales of pharmaceutical products and related activities, such as arrangements with investigators, healthcare professionals, consultants, third-party payors, patient organizations, and customers, are subject to fraud and abuse and other healthcare laws and regulations, which are enforced by the federal government and the states and foreign governments in which the business is conducted. Applicable healthcare laws and regulations that may affect a company’s ability to operate if and when marketing approval is granted for a product candidate include the following:
| ∎ | the federal Anti-Kickback Statute, which prohibits, among other things, individuals and entities from knowingly and willfully soliciting, receiving, offering, or providing remuneration (including any kickback, bribe or rebate), directly or indirectly, overtly or covertly, in cash or in kind, to induce or reward, or in return for, either the referral of an individual for, or the purchase, order, or recommendation of, any good or service for which payment may be made, in whole or in part, under a federal or state healthcare program, such as Medicare or Medicaid. The term “remuneration” has been broadly interpreted to include anything of value. Rather, if “one purpose” of the remuneration is to induce referrals, the federal Anti-Kickback Statute is violated. In addition, a person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation. Violations are subject to civil and criminal fines and penalties for each violation, plus up to three times the remuneration involved, imprisonment, and exclusion from federal programs; |
| ∎ | the federal criminal and civil false claims laws, including the federal False Claims Act (FCA), which can be enforced through civil whistleblower or “qui tam” actions against individuals or entities, and prohibits, among other things, knowingly presenting, or causing to be presented to the federal government claims for payment that are false or fraudulent, knowingly making, using or causing to be made or used, a false record or statement material to a false or fraudulent claim, or from knowingly making a false statement to avoid, decrease or conceal an obligation to pay money to the federal government. Manufacturers can be held liable under the FCA even when they do not submit claims directly to government payors if they are deemed to “cause” the submission of false or fraudulent claims. In addition, certain marketing practices, including off-label promotion, may also violate false claims laws. Moreover, the government may assert that a claim including items and services resulting from a violation of the federal Anti-Kickback Statute constitutes a false or fraudulent claim for purposes of the FCA. The FCA also permits a private individual acting as a “whistleblower” to bring actions on behalf of the federal government alleging violations of the FCA and to share in any monetary recovery; |
| ∎ | the Health Insurance Portability and Accountability Act (HIPAA), which prohibits, among other things, knowingly and willfully executing, or attempting to execute, a scheme to defraud any healthcare benefit |
143
Table of Contents
| program or obtain, by means of false or fraudulent pretenses, representations or promises, any of the money or property owned by, or under the custody or control of, any healthcare benefit program, regardless of payor (e.g., public or private), or knowingly and willfully falsifying, concealing or covering up a material fact or making any materially false statement in connection with the delivery of or payment for healthcare benefits, items or services. Similar to the Anti-Kickback Statute, a person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation; |
| ∎ | HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act of 2009 (HITECH) and their respective implementing regulations, which impose obligations on certain covered healthcare providers, health plans, and healthcare clearinghouses, as well as their respective business associates and subcontractors that perform certain services involving the storage, use or disclosure of individually identifiable health information for or on behalf of a covered entity and their business associates, including mandatory contractual terms, with respect to safeguarding the privacy, security, and transmission of individually identifiable health information, and require notification to affected individuals and regulatory authorities of certain breaches of security of individually identifiable health information. HITECH also created new tiers of civil monetary penalties, amended HIPAA to make civil and criminal penalties directly applicable to business associates, and gave state attorneys general new authority to file civil actions for damages or injunctions in federal courts to enforce HIPAA and seek attorneys’ fees and costs associated with pursuing federal civil actions; |
| ∎ | the federal Physician Payments Sunshine Act, which requires certain manufacturers of drugs, devices, biologics, and medical supplies for which payment is available under Medicare, Medicaid, or the Children’s Health Insurance Program, with certain exceptions, to report annually to the Centers for Medicare & Medicaid Services (CMS) information related to payments or other transfers of value made to physicians (defined to include doctors, dentists, optometrists, podiatrists, and chiropractors), certain other health care professionals (such as physician assistants, nurse practitioners, clinical nurse specialists, anesthesiologist assistants, certified registered nurse anesthetists, and certain nurse midwives) and teaching hospitals, as well as ownership and investment interests held by the physicians described above and their immediate family members, with the information made publicly available on a searchable website; |
| ∎ | analogous state and foreign laws and regulations, such as state anti-kickback and false claims laws, that may apply to sales or marketing arrangements and claims involving healthcare items or services reimbursed under Medicaid and other state programs, or in several states, apply regardless of payor, including private insurers and cash-pay patients; |
| ∎ | state laws that require the registration of manufacturers and wholesale distributors of drug and biological products who ship into a state, including in certain states that require registration even if such manufacturers or distributors have no place of business within the state. Some states also impose requirements on manufacturers and distributors to establish the pedigree of product in the chain of distribution, including some states that require manufacturers and others to adopt new technology capable of tracking and tracing product as it moves through the distribution chain; |
| ∎ | certain state laws that require pharmaceutical and biotechnology companies to establish marketing compliance programs and comply with the pharmaceutical industry’s voluntary compliance guidelines and the relevant compliance guidance promulgated by the federal government in addition to requiring drug manufacturers to report information related to payments to physicians and other healthcare or marketing expenditures and drug pricing information, and state and local laws that require the registration of pharmaceutical sales representatives, and to prohibit certain other sales and marketing practices; |
| ∎ | analogous foreign laws and regulations, including restrictions imposed on the promotion and marketing of medicinal products in European Union (EU) member states and other countries, restrictions on interactions with healthcare professionals and requirements for public disclosure of payments made to physicians. Laws (including those governing promotion, marketing and anti-kickback provisions), industry regulations, and professional codes of conduct often are strictly enforced. |
Additionally, we are subject to foreign as well as U.S. federal and state laws and regulations governing the collection, use, access to, confidentiality, privacy and security of health-related and other personal information, many of which differ from each other in significant ways and often are not preempted by HIPAA. Violations of any such requirements, may result in significant penalties, including civil, criminal and administrative penalties, damages,
144
Table of Contents
fines, disgorgement, imprisonment, the curtailment or restructuring of operations, loss of eligibility to obtain approvals from the FDA or foreign regulatory authorities, exclusion from participation in government contracting, healthcare reimbursement or other government programs, including Medicare and Medicaid, integrity oversight and reporting obligations, or reputational harm.
Pharmaceutical Coverage, Pricing and Reimbursement
Significant uncertainty exists as to the coverage and reimbursement status of any product candidates for which we obtain regulatory approval. In the United States and markets in other countries, sales of any products for which we receive regulatory approval for commercial sale will depend, in part, on the extent to which third-party payors provide coverage, and establish adequate reimbursement levels for such products. In the United States, third-party payors include federal and state healthcare programs, government authorities, private managed care providers, private health insurers, and other organizations, where there is no uniform policy for coverage and reimbursement and can differ significantly from payor to payor.
Third-party payors decide which drugs and treatments they will cover and the amount of reimbursement. Reimbursement by a third-party payor may depend upon a number of factors, including, but not limited to, the third-party payor’s determination that use of a product is:
| ∎ | a covered benefit under its health plan; |
| ∎ | safe, effective and medically necessary; |
| ∎ | appropriate for the specific patient; |
| ∎ | cost-effective; and |
| ∎ | neither experimental nor investigational. |
Third-party payors are increasingly challenging the price, examining the medical necessity and reviewing the cost-effectiveness of medical drug products and medical services, in addition to questioning their safety and efficacy. Such payors may limit coverage to specific drug products on an approved list, also known as a formulary, which might not include all of the FDA-approved drugs for a particular indication. Companies may need to conduct expensive pharmaco-economic studies in order to demonstrate the medical necessity and cost-effectiveness of their products, in addition to the costs required to obtain the FDA approvals. Nonetheless, a product candidate may not be considered medically necessary or cost-effective. Moreover, the process for determining whether a third-party payor will provide coverage for a drug product may be separate from the process for setting the price of a drug product or for establishing the reimbursement rate that such a payor will pay for the drug product. A payor’s decision to provide coverage for a drug product does not imply that an adequate reimbursement rate will be approved. Further, one payor’s determination to provide coverage for a drug product does not assure that other payors will also provide coverage for the drug product.
Governments and private third-party payors have attempted to control costs by limiting coverage and the amount of reimbursement for particular medications. For example, the U.S. Department of Health and Human Services (HHS) imposes rebates on many Medicare Part B and Medicare Part D products to penalize price increases that outpace inflation. HHS has also been empowered to negotiate the price of certain single-source drugs that have been on the market for at least seven (7) years without being the listed product for a generic, or certain BLA-licensed biological products on the market for at least eleven (11) years without being the reference biological product for a biosimilar, covered under Medicare as part of the Medicare drug price negotiation program. Each year, up to twenty (20) products will be selected by HHS for the Medicare drug price negotiation program. Products subject to the Medicare drug price negotiation program are expected to experience a significant reduction in reimbursement from the Medicare program on a per unit basis. Adequate third-party reimbursement may not be available to enable a company to maintain price levels sufficient to realize an appropriate return on its investment in product development.
Outside the United States, pricing of prescription pharmaceuticals is subject to governmental control in many countries. Pricing negotiations with governmental authorities can extend well beyond the receipt of regulatory marketing approval for a product. In the EU, pricing and reimbursement schemes vary widely from one member state to another. Some member states may require the completion of additional studies that compare the cost-effectiveness of a particular medicinal product candidate to currently available therapies or so-called Health
145
Table of Contents
Technology Assessments (HTA), in order to obtain reimbursement or pricing approval. For example, the EU provides options for its member states to restrict the range of products for which their national health insurance systems provide reimbursement and to control the prices of medicinal products for human use. EU member states may approve a specific price for a product or it may instead adopt a system of direct or indirect controls on the profitability of the company placing the product on the market. Other EU member states allow companies to fix their own prices for products, but monitor and control prescription volumes and issue guidance to physicians to limit prescriptions. The downward pressure on healthcare costs in general, and particularly in relation to prescription only medicinal products, has become more intense. As a result, increasingly high barriers are being erected to the entry of new products.
The marketability of any product candidates for which regulatory approval is granted for commercial sale may suffer if the government and third-party payors fail to provide adequate coverage and reimbursement. In addition, emphasis on managed care in the United States has increased and could increase the pressure on pharmaceutical pricing. Coverage policies and third-party reimbursement rates may change at any time. Even if favorable coverage and reimbursement status is attained for one or more products for which regulatory approval is granted, less favorable coverage policies and reimbursement rates may be implemented in the future.
Healthcare Reform
The United States and many foreign jurisdictions have enacted or proposed legislative and regulatory changes affecting the healthcare system. The United States government, state legislatures, and foreign governments also have shown significant interest in implementing cost-containment programs to limit the growth of government-paid healthcare costs, including price controls, restrictions on reimbursement, and requirements for substitution of generic products for branded prescription drugs and biologics. In recent years, Congress has considered reductions in Medicare reimbursement levels for drugs and biologics administered by physicians. CMS, the agency that administers the Medicare and Medicaid programs, also has authority to revise reimbursement rates and to implement coverage restrictions for some drugs and biologics. Cost reduction initiatives and changes in coverage implemented through legislation or regulation could decrease utilization of and reimbursement for any approved products. While Medicare regulations apply only to drug benefits for Medicare beneficiaries, private payors often follow Medicare coverage policy and payment limitations in setting their own reimbursement rates. Therefore, any reduction in reimbursement that results from federal legislation or regulation may result in a similar reduction in payments from private payors.
Additionally, in March 2010, the Patient Protection and Affordable Care Act, as amended by the Health Care and Education Reconciliation Act (collectively, the ACA) was enacted, which substantially changed the way healthcare is financed by both governmental and private insurers. Since its enactment, there have been amendments and judicial, Congressional and executive branch challenges to certain aspects of the ACA. For example, on July 4, 2025, the One Big Beautiful Bill Act (the OBBBA) was signed into law, which narrowed access to ACA marketplace exchange enrollment and declined to extend the ACA enhanced advanced premium tax credits that expired at the end of 2025, which, among other provisions in the law, are anticipated to reduce the number of Americans with health insurance. The OBBBA also is expected to reduce Medicaid spending and enrollment by implementing work requirements for some beneficiaries, capping state-directed payments, reducing federal funding, and limiting provider taxes used to fund the program. Congress is considering proposed legislation intended to further reduce healthcare costs with alternatives to replace the expired ACA subsidies. We expect that additional U.S. federal healthcare reform measures will be adopted in the future, any of which could limit the amounts that the U.S. federal government will pay for healthcare products and services, which could result in reduced demand for our product candidates or additional pricing pressures.
Other legislative changes have been proposed and adopted in the United States since the ACA was enacted. These changes include aggregate reductions to Medicare payments to providers, which began in 2013 and will remain in effect until 2032 unless additional Congressional action is taken.
The federal administration is pursuing policies to reduce regulations and expenditures across government, including at HHS, which include the FDA and CMS, and related agencies. These actions, presently directed by executive orders or memoranda from the Office of Management and Budget, may propose policy changes that create additional uncertainty for our business. For example, the current administration has announced agreements with several
146
Table of Contents
pharmaceutical companies that require the drug manufacturers to offer, through a direct to consumer platform (TrumpRx) U.S. patients and Medicaid programs prescription drug Most-Favored Nation pricing equal to or lower than those paid in other developed nations, with additional mandates for direct-to-patient discounts and repatriation of foreign revenues. Other recent actions, for example, include (1) directing agencies to reduce agency workforce and cut programs; (2) directing HHS and other agencies to lower prescription drug costs through a variety of initiatives; (3) imposing tariffs on certain imported pharmaceutical products; and (4) as part of the Make America Healthy Again (MAHA) Commission’s Strategy Report released in September 2025, working across government agencies to increase enforcement on direct-to-consumer pharmaceutical advertising. Additionally, the current administration recently called on Congress to enact “The Great Healthcare Plan,” to codify and expand Most-Favored Nation pricing, lower government subsidies to private insurance companies, increase healthcare price transparency, expand pharmaceutical drugs available for over-the-counter purchase, and enact restrictions on pharmacy benefit manager payment methodologies, among other things. Further, in June 2024, the U.S. Supreme Court’s Loper Bright decision greatly reduced judicial deference to regulatory agencies, which could increase successful legal challenges to federal regulations affecting our operations. These actions and policies may significantly reduce U.S. drug prices, potentially impacting manufacturers’ global pricing strategies and profitability, while increasing their operational costs and compliance risks.
At the state level, legislatures have increasingly passed legislation and implemented regulations designed to control pharmaceutical product pricing, including price or patient reimbursement constraints, discounts, restrictions on certain product access, and marketing cost disclosure and transparency measures, and in some cases, designed to encourage importation from other countries and bulk purchasing.
Additional federal, state, and foreign healthcare reform measures may be adopted in the future, any of which could limit the amounts that federal and state governments will pay for healthcare products and services, which could result in limited coverage and reimbursement and reduced demand for pharmaceutical products or additional pricing pressures.
For instance, in the EU, on December 15, 2021, Regulation No 2021/2282 on HTA, amending Directive 2011/24/EU, was adopted. The Regulation entered into force in January 2022 and has been applicable since January 2025, with phased implementation based on the type of product, i.e. oncology and advanced therapy medicinal products as of 2025, orphan medicinal products as of 2028, and all other medicinal products by 2030. The Regulation intends to boost cooperation among EU member states in assessing health technologies, including new medicinal products, and provide the basis for cooperation at the EU level for joint clinical assessments in these areas. It will permit EU member states to use common HTA tools, methodologies, and procedures across the EU, working together in four main areas, including joint clinical assessment of the innovative health technologies with the highest potential impact for patients, joint scientific consultations whereby developers can seek advice from HTA authorities, identification of emerging health technologies to identify promising technologies early, and continuing voluntary cooperation in other areas. Individual EU member states will continue to be responsible for assessing non-clinical (e.g., economic, social, ethical) aspects of health technology, and making decisions on pricing and reimbursement.
On December 11, 2025, the European Commission, the Parliament and the European Council reached a political agreement on a comprehensive overhaul of EU pharmaceutical legislation (the Pharma Package). The reform has been under negotiation since the European Commission submitted its proposal in April 2023. This package—comprised of a new directive and regulation to replace existing legislation—aims to modernize the EU regulatory framework. The political agreement is still subject to formal approval by the European Parliament and Council. If approved in the form proposed, the Pharma Package will, among other changes, reduce the baseline market protection period by one year, with limited opportunities for extensions; reshape the incentives regime for orphan medicinal products; and expand the Bolar exemption. A decrease in market exclusivity opportunities for our product candidates in the EU, combined with the expanded Bolar exemption, could open them to generic or biosimilar competition earlier than under the current regime, potentially impacting reimbursement status and the commercial prospects of our product candidates. The new framework is expected to enter into force in 2026/2027 and to be subject to transitional arrangements, with full application not anticipated before 2028.
147
Table of Contents
Foreign Government Regulation
To market any product outside of the United States, we would need to comply with numerous and varying regulatory requirements of other countries governing, among other things, clinical trials, marketing authorization (MA), commercial sales and distribution of our products.
Whether or not we obtain FDA approval for a product, we must obtain approval of a product by the comparable regulatory authorities of foreign countries before we can commence clinical trials or marketing of the product in those countries. Approval by one regulatory authority does not ensure approval by regulatory authorities in other jurisdictions. The approval process varies from country to country, can involve additional testing beyond that required by FDA, and may be longer or shorter than that required for FDA approval. The requirements governing the conduct of clinical trials, product licensing, pricing, promotion, and reimbursement vary greatly from country to country.
Non-clinical Studies and Clinical Trials
Similarly to the United States, the various phases of non-clinical and clinical research in the EU are subject to significant regulatory controls.
Non-clinical studies are performed to demonstrate the health or environmental safety of new chemical or biological substances. Non-clinical (pharmaco-toxicological) studies must be conducted in compliance with the principles of GLP, as set forth in EU Directive 2004/10/EC (unless otherwise justified for certain particular medicinal products, e.g., radio-pharmaceutical precursors for radio-labeling purposes). In particular, non-clinical studies, both in vitro and in vivo, must be planned, performed, monitored, recorded, reported, and archived in accordance with the GLP principles, which define a set of rules and criteria for a quality system for the organizational process and the conditions for non-clinical studies. These GLP standards reflect the Organization for Economic Co-operation and Development requirements.
Clinical trials of medicinal products in the EU must be conducted in accordance with EU and national regulations and the International Council for Harmonization of Technical Requirements for Pharmaceuticals for Human Use (ICH) guidelines on GCP as well as the applicable regulatory requirements and the ethical principles that have their origin in the Declaration of Helsinki. If the sponsor of the clinical trial is not established within the EU, it must appoint an EU entity to act as its legal representative. The sponsor must take out a clinical trial insurance policy, and in most EU member states, the sponsor is liable to provide ‘no fault’ compensation to any study subject injured in the clinical trial.
The regulatory landscape related to clinical trials in the EU has been subject to recent changes. The EU Clinical Trials Regulation (CTR) which was adopted in April 2014 and repeals the EU Clinical Trials Directive, became applicable on January 31, 2022. Unlike directives, the CTR is directly applicable in all EU member states without the need for member states to further implement it into national law. The CTR notably harmonizes the assessment and supervision processes for clinical trials throughout the EU via a Clinical Trials Information System, which contains a centralized EU portal and database.
While the EU Clinical Trials Directive required a separate clinical trial application (CTA) to be submitted in each member state in which the clinical trial takes place, to both the competent national health authority and an independent ethics committee, much like the FDA and IRB respectively, the CTR introduces a centralized process and only requires the submission of a single application for multi-center trials. The CTR allows sponsors to make a single submission to both the competent authority and an ethics committee in each member state, leading to a single decision per member state. The CTA must include, among other things, a copy of the trial protocol and an investigational medicinal product dossier containing information about the manufacture and quality of the medicinal product under investigation. The assessment procedure of the CTA has been harmonized as well, including a joint assessment by all member states concerned, and a separate assessment by each member state with respect to specific requirements related to its own territory, including ethics rules. Each member state’s decision is communicated to the sponsor via the centralized EU portal. Once the CTA is approved, clinical study development may proceed.
The CTR transition period ended on January 31, 2025, and all clinical trials (and related applications) are now fully subject to the provisions of the CTR.
148
Table of Contents
Medicines used in clinical trials must be manufactured in accordance with Good Manufacturing Practice (GMP). Other national and EU-wide regulatory requirements may also apply.
Marketing Authorization
In order to market our product candidates in the EU and many other foreign jurisdictions, we must obtain separate regulatory approvals. More concretely, in the EU, medicinal product candidates can only be commercialized after obtaining an MA. To obtain regulatory approval of a product candidate under EU regulatory systems, we must submit an MA application (MAA). The process for doing this depends, among other things, on the nature of the medicinal product. There are two types of MAs:
| ∎ | “Centralized MAs” are issued by the European Commission through the centralized procedure based on the opinion of the Committee for Medicinal Products for Human Use (CHMP), of the European Medicines Agency (EMA), and are valid throughout the EU. The centralized procedure is compulsory for certain types of medicinal products such as (i) medicinal products derived from biotechnological processes, (ii) designated orphan medicinal products, (iii) advanced therapy medicinal products (ATMPs) (such as gene therapy, somatic cell therapy and tissue engineered products), and (iv) medicinal products containing a new active substance indicated for the treatment of certain diseases, such as HIV/AIDS, cancer, diabetes, neurodegenerative diseases or autoimmune diseases and other immune dysfunctions, and viral diseases. The centralized procedure is optional for products containing a new active substance not yet authorized in the EU, or for products that constitute a significant therapeutic, scientific or technical innovation or which are in the interest of public health in the EU. |
| ∎ | “National MAs” are issued by the competent authorities of the EU member states, only cover their respective territory, and are available for product candidates not falling within the mandatory scope of the centralized procedure. Where a product has already been authorized for marketing in an EU member state, this national MA can be recognized in another member state through the mutual recognition procedure. If the product has not received a national MA in any member state at the time of application, it can be approved simultaneously in various member states through the decentralized procedure. Under the decentralized procedure an identical dossier is submitted to the competent authorities of each of the member states in which the MA is sought, one of which is selected by the applicant as the reference member state. |
Under the centralized procedure the maximum timeframe for the evaluation of an MAA by the EMA is 210 days, excluding clock stops. In exceptional cases, the CHMP might perform an accelerated review of an MAA in no more than 150 days (not including clock stops).
Innovative products that target an unmet medical need and are expected to be of major public health interest may be eligible for a number of expedited development and review programs, such as the PRIME scheme, which provides incentives similar to the breakthrough therapy designation in the U.S. In March 2016, the EMA launched an initiative, the PRIME scheme, a voluntary scheme aimed at enhancing the EMA’s support for the development of medicines that target unmet medical needs. It is based on increased interaction and early dialogue with companies developing promising medicines, to optimize their product development plans and speed up their evaluation to help them reach patients earlier. In July 2025, ipsoprubart was granted PRIME designation by the EMA. Product developers that benefit from PRIME designation can expect to be eligible for accelerated assessment but this is not guaranteed. Many benefits accrue to sponsors of product candidates with PRIME designation, including but not limited to, early and proactive regulatory dialogue with the EMA, frequent discussions on clinical trial designs and other development program elements, and accelerated MAA assessment once a dossier has been submitted. Importantly, a dedicated contact and rapporteur from the CHMP is appointed early in the PRIME scheme facilitating increased understanding of the product at EMA’s committee level. An initial meeting initiates these relationships and includes a team of multidisciplinary experts at the EMA to provide guidance on the overall development and regulatory strategies.
Under the above described procedures, in order to grant the MA, the EMA or the competent authorities of the EU member states make an assessment of the risk benefit balance of the product on the basis of scientific criteria concerning its quality, safety, and efficacy. MAs have an initial duration of five years. After these five years, the authorization may be renewed on the basis of a reevaluation of the risk-benefit balance.
149
Table of Contents
Data and Marketing Exclusivity
In the EU, new products authorized for marketing (i.e., reference products) generally receive eight years of data exclusivity and an additional two years of market exclusivity upon MA. If granted, the data exclusivity period prevents generic and biosimilar applicants from relying on the preclinical and clinical trial data contained in the dossier of the reference product when applying for a generic or biosimilar MA in the EU during a period of eight years from the date on which the reference product was first authorized in the EU. The market exclusivity period prevents a successful generic or biosimilar applicant from commercializing its product in the EU until ten years have elapsed from the initial MA of the reference product in the EU. The overall ten-year market exclusivity period can be extended to a maximum of eleven years if, during the first eight years of those ten years, the MA holder obtains an authorization for one or more new therapeutic indications, which, during the scientific evaluation prior to their authorization, are held to bring a significant clinical benefit in comparison with existing therapies. However, there is no guarantee that a product will be considered by the EU’s regulatory authorities to be a new chemical or biological entity, and products may not qualify for data exclusivity.
There is a special regime for biosimilars, or biological medicinal products that are similar to a reference medicinal product but that do not meet the definition of a generic medicinal product, for example, because of differences in raw materials or manufacturing processes. For such products, the results of appropriate preclinical or clinical trials must be provided, and guidelines from the EMA detail the type of quantity of supplementary data to be provided for different types of biological product. There are no such guidelines for complex biological products, such as gene or cell therapy medicinal products, and so it is unlikely that biosimilars of those products will currently be approved in the EU. However, guidance from the EMA states that they will be considered in the future in light of the scientific knowledge and regulatory experience gained at the time.
Orphan Medicinal Products
The criteria for designating an “orphan medicinal product” in the EU are similar in principle to those in the United States. A medicinal product can be designated as an orphan if its sponsor can establish that: (1) the product is intended for the diagnosis, prevention or treatment of a life threatening or chronically debilitating condition; (2) either (a) such condition affects not more than five in 10,000 persons in the EU when the application is made, or (b) the product, without the benefits derived from the orphan status, would not generate sufficient return in the EU to justify the necessary investment; and (3) there exists no satisfactory method of diagnosis, prevention or treatment of the condition in question that has been authorized for marketing in the EU or, if such method exists, the product will be of significant benefit to those affected by that condition.
Orphan designation must be requested before submitting an MAA. An EU orphan designation entitles a party to incentives such as reduction of fees or fee waivers, protocol assistance, and access to the centralized procedure. Upon grant of a MA, orphan medicinal products are entitled to ten years of market exclusivity for the approved indication, which means that the competent authorities cannot accept another MAA, or grant a MA, or accept an application to extend a MA for a similar medicinal product for the same indication for a period of ten years. The period of market exclusivity is extended by two years for orphan medicinal products that have also complied with an agreed pediatric investigation plan (PIP). No extension to any supplementary protection certificate can be granted on the basis of pediatric studies for orphan indications. Orphan designation does not convey any advantage in, or shorten the duration of, the regulatory review and approval process.
The orphan exclusivity period may be reduced to six years if, at the end of the fifth year, it is established that the product no longer meets the criteria for which it received orphan designation, including where it is shown that the product is sufficiently profitable not to justify maintenance of market exclusivity or where the prevalence of the condition has increased above the threshold. Additionally, MA may be granted to a similar product for the same indication at any time if (i) the second applicant can establish that its product, although similar, is safer, more effective or otherwise clinically superior; (ii) the applicant consents to a second orphan medicinal product application; or (iii) the applicant cannot supply enough orphan medicinal product.
Pediatric Development
In the EU, MAAs for new medicinal products candidates have to include the results of studies conducted in the pediatric population, in compliance with a PIP agreed with the EMA’s Pediatric Committee (PDCO). The PIP sets out the timing and measures proposed to generate data to support a pediatric indication of the product candidate for which MA is being sought. The PDCO can grant a deferral of the obligation to implement some or all of the measures
150
Table of Contents
of the PIP until there are sufficient data to demonstrate the efficacy and safety of the product in adults. Further, the obligation to provide pediatric clinical trial data can be waived by the PDCO when these data are not needed or appropriate because the product is likely to be ineffective or unsafe in children, the disease or condition for which the product is intended occurs only in adult populations, or when the product does not represent a significant therapeutic benefit over existing treatments for pediatric patients. Once the MA is obtained in all the EU member states and study results are included in the product information, even when negative, the product is eligible for six months’ supplementary protection certificate extension (if any is in effect at the time of approval) or, in the case of orphan medicinal products, a two year extension of the orphan market exclusivity is granted.
The aforementioned EU rules are generally applicable in the European Economic Area (EEA) which consists of the 27 EU member states plus Norway, Liechtenstein and Iceland.
Failure to comply with applicable foreign regulatory requirements, including EU and member state laws, may result in administrative, civil or criminal penalties. These penalties could include delays or refusal to authorize the conduct of clinical trials, or to grant MA, product withdrawals and recalls, product seizures, suspension, withdrawal or variation of the MA, total or partial suspension of production, distribution, manufacturing or clinical trials, operating restrictions, injunctions, suspension of licenses, fines and criminal penalties.
EU Pharmaceutical Reform
On December 11, 2025, the European Commission, the Parliament and the European Council reached a political agreement on a comprehensive overhaul of EU pharmaceutical legislation (the Pharma Package). The reform has been under negotiation since the European Commission submitted its proposal in April 2023. This package—comprised of a new directive and regulation to replace existing legislation—aims to modernize the EU regulatory framework. The political agreement is still subject to formal approval by the European Parliament and Council. If approved in the form proposed, the Pharma Package will, among other changes, reduce the baseline market protection period by one year, with limited opportunities for extensions; reshape the incentives regime for orphan medicinal products; and expand the Bolar exemption. A decrease in market exclusivity opportunities for our product candidates in the EU, combined with the expanded Bolar exemption, could open them to generic or biosimilar competition earlier than under the current regime, potentially impacting reimbursement status and the commercial prospects of our product candidates. The new framework is expected to enter into force in 2026/2027 and to be subject to transitional arrangements, with full application not anticipated before 2028.
The Foreign Corrupt Practices Act
The Foreign Corrupt Practices Act of 1977, as amended (FCPA), prohibits any U.S. individual or business from paying, offering, or authorizing payment or offering of anything of value, directly or indirectly, to any foreign official, political party or candidate for the purpose of influencing any act or decision of the foreign entity in order to assist the individual or business in obtaining or retaining business. U.S. authorities that enforce the FCPA, including the DOJ, deem most health care professionals and other employees of foreign hospitals, clinics, research facilities and medical schools in countries with public health care or public education systems to be “foreign officials” under the FCPA. The FCPA also obligates companies whose securities are listed in the United States to comply with accounting provisions requiring the company to maintain books and records that accurately and fairly reflect all transactions of the corporation, including international subsidiaries and to devise and maintain an adequate system of internal accounting controls for international operations.
Data Privacy and Security
Numerous state, federal, and foreign laws, regulations and standards govern the collection, use, access to, confidentiality, and security of personal information and health-related data, including clinical trial data, and could apply now or in the future to our operations or the operations of our partners. In the United States, numerous federal and state laws and regulations, including data breach notification laws, health information privacy and security laws, comprehensive data privacy laws, and consumer protection laws and regulations, govern the collection, use, disclosure, and protection of personal information, including health-related data. Further, our collection and processing of personal information from individuals in the EU and other jurisdictions outside of the United States, through clinical trials or otherwise, may render us subject to foreign laws, such as the GDPR, which govern the privacy and security of personal information, including health-related data. Our use of machine learning technology
151
Table of Contents
may also be subject to evolving laws and regulations, which may require us to, among other things, make specific disclosures or control unwanted bias and discrimination. Privacy and security laws, regulations and other obligations are constantly evolving, may conflict with each other to complicate compliance efforts, and can result in investigations, proceedings, or actions that lead to significant civil and/or criminal penalties and restrictions on data processing.
Employees and Human Capital Resources
As of June 30, 2026, we had 46 employees, all of whom were full-time and 33 of whom were engaged in research and development activities. Approximately 24 percent of our employees hold Ph.D., Pharm.D. or M.D. or other advanced degrees. None of our employees are represented by a labor union or covered under a collective bargaining agreement. We consider our relationship with our employees to be good.
Our human capital resources objectives include, as applicable, identifying, recruiting, retaining, incentivizing, and integrating our existing and new employees, advisors and consultants. It is important that we not only attract and retain the best and brightest diverse talent, but also ensure they remain engaged and can thrive in an environment that is committed to helping them grow, succeed and contribute directly to achieving our purpose. The principal purposes of our equity and cash incentive plans are to attract, retain and reward personnel through the granting of stock-based and cash-based compensation awards, in order to increase the success of our Company by motivating such individuals to perform to the best of their abilities and achieve our objectives. We also strive to foster career growth and internal mobility by providing a broad range of training, mentoring and other development opportunities.
Facilities
Our headquarters are located in South San Francisco, California where we lease and occupy approximately 14,055 square feet of office and laboratory space. The current term of our lease expires in September 2028. We have the option to extend the lease for an additional three years after initial expiration. We believe that our existing facilities are sufficient to meet our near-term needs and that suitable additional space will be available as and when needed.
Legal Proceedings
From time to time, we may be subject to legal proceedings. We are not currently a party to or aware of any legal proceedings that we believe will have, individually or in the aggregate, a material adverse effect on our business, financial condition or results of operations. Regardless of outcome, litigation can have an adverse impact on us because of defense and settlement costs, diversion of management resources, and other factors.
Corporate Information
We were originally incorporated under the laws of the State of Delaware in October 2018 as a wholly-owned subsidiary of Star Therapeutics LLC (Star), a private biotechnology company. In February 2022, in connection with our Series B Preferred Stock financing, we became a majority-owned subsidiary of Star. In June 2023, Star formed Electra Therapeutics LLC (Electra LLC) and contributed all of the equity it held in us to Electra LLC in exchange for the equity of Electra LLC, which Star subsequently distributed to its equity holders (the Spin-Out). As a result of the Spin-Out, Star is no longer our parent company and we are no longer directly affiliated with Star.
152
Table of Contents
The following table sets forth certain information for our executive officers and directors as of August 1, 2026:
| NAME |
AGE |
POSITION(S) | ||
| Executive Officers | ||||
| Quehuong (Kathy) Dong, Pharm.D., M.B.A. |
44 | President, Chief Executive Officer, Director | ||
| Chris Clark, C.F.A. |
48 | Chief Financial Officer | ||
| Kim-Hien Dao, D.O., Ph.D. |
53 | Chief Medical Officer | ||
| Gary S. Koe, Ph.D. |
62 | Chief Technical Officer | ||
| Graham Parry, Ph.D. |
62 | Chief Scientific Officer | ||
| Non-Employee Directors | ||||
| Nancy Stagliano, Ph.D.(1)(3) |
59 | Chairperson | ||
| Matthew Fust, M.B.A.(1)(2)(3) |
62 | Director | ||
| Thomas Geninatti, Ph.D.(4) |
39 | Director | ||
| Carl L. Gordon, Ph.D., C.F.A.(1)(2) |
61 | Director | ||
| Beth Seidenberg, M.D.(4) |
69 | Director | ||
| (1) | Member of the audit committee. |
| (2) | Member of the compensation committee. |
| (3) | Member of the nominating and corporate governance committee. |
| (4) | Drs. Geninatti and Seidenberg have notified us that they intend to resign from our board of directors effective as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. |
Executive Officers
Quehuong (Kathy) Dong, Pharm.D., M.B.A., has served as our President and Chief Executive Officer and a member of our board of directors since October 2023. She has also served as President and Chief Executive Officer and a member of the board of managers of Electra Therapeutics LLC since October 2023. From April 2019 to October 2023, Dr. Dong served as Chief Operating Officer of Star Therapeutics, Inc., a privately held biotechnology company (Star Therapeutics), where she held strategic and operational responsibilities across the company’s portfolio. Dr. Dong also served as Chief Operating Officer at Vega Therapeutics Inc., a privately held biotechnology company, from April 2019 to October 2023. Prior to Star Therapeutics, Dr. Dong served as Vice President of Commercial Development at True North Therapeutics, Inc., a privately held biotechnology company (True North). Following True North’s acquisition by Bioverativ, Inc. (which was subsequently acquired by Sanofi S.A. (Sanofi)), Dr. Dong served as Head of New Product Planning for the complement franchise at Sanofi, a publicly held pharmaceutical company. Her tenure from True North through Sanofi was March 2017 to March 2019. Earlier in her career, Dr. Dong served as Director, HCV Marketing at Gilead Sciences, Inc., a publicly held biopharmaceutical company (Gilead). Dr. Dong has also served on the board of directors of Neuron23, Inc., a privately held precision neurology and immunology therapeutics company (Neuron23), since June 2022. Dr. Dong earned her Pharm.D. and M.B.A. at the University of New Mexico and completed a postdoctoral fellowship through Rutgers University.
Our board of directors believes that Dr. Dong’s distinguished scientific background and experience in leadership roles in the biopharmaceutical industry, as well as the perspective and experience she brings as our President and Chief Executive Officer, qualifies her to serve on our board of directors.
Chris Clark, C.F.A., has served as our Chief Financial Officer since August 2026 and as our Executive Vice President, Strategy and Finance from December 2025 to August 2026. Previously, Mr. Clark served as Portfolio Manager, Equity Analyst – HealthCare at RS Investments, a Victory Capital investment franchise, a publicly held investment management company, from May 2007 to May 2025. Mr. Clark also served as a Research Associate at TIAA-CREF, a financial services organization, from 2004 to 2007, where he focused on global pharmaceuticals. Mr. Clark earned his B.A. in economics from the University of Virginia and is a chartered financial analyst.
Kim-Hien Dao, D.O., Ph.D., has served as our Chief Medical Officer since May 2024 and previously served as our Vice President, Head of Clinical Development from April 2023 to May 2024. Prior to joining us, Dr. Dao served as
153
Table of Contents
Senior and Executive Medical Director of Clinical Development at Astex Pharmaceuticals, Inc., a privately held pharmaceutical company, from November 2019 to March 2023, where she worked on drug development programs for solid tumors and hematological malignancies. Dr. Dao earned her D.O. and Ph.D. in biochemistry and molecular biology from Michigan State University and trained in hematology-medical oncology at the University of California, San Diego.
Gary S. Koe, Ph.D., has served as our Chief Technical Officer since May 2025. Previously, Dr. Koe served as our Senior Vice President, Head of Technical Operations from May 2024 to May 2025 and Vice President, Head of Technical Operations from November 2023 to May 2024. Dr. Koe was also Vice President, Technical Operations at Star Therapeutics from March 2022 to November 2023. Dr. Koe also served as Executive Director, Head of Operations at Star Therapeutics from November 2019 to March 2022. Prior to joining us, Dr. Koe served as Executive Director of Technical Operations at Aravive, Inc., a biotechnology company (Aravive), from 2015 to 2019. Dr. Koe earned a Ph.D. in chemical engineering from the University of California, Los Angeles, and his B.S. in chemical engineering from the University of California, Davis.
Graham Parry, Ph.D., has served as our Chief Scientific Officer since December 2023. Prior to joining us, Dr. Parry served as Executive Vice President, Translational Sciences at Star Therapeutics from January 2023 to December 2023, where he worked cross-functionally across the company’s portfolio to advance treatments for multiple diseases. Prior to Star Therapeutics, Dr. Parry served as Senior Vice President, Translational Sciences at Third Harmonic Bio, Inc., a privately held clinical-stage biotechnology company, from August 2019 to December 2022. Dr. Parry earned a Ph.D. in biochemistry at the University of London and a B.Sc. in applied biology at Brunel University of London.
Non-Employee Directors
Nancy Stagliano, Ph.D., has served as our Chairperson and member of our board of directors since March 2019. Dr. Stagliano has served as the chair of the board of directors of Neuron23 since October 2018 and as Chief Executive Officer of Neuron23 since October 2020. Dr. Stagliano has also served as chair of the board of directors of Star Therapeutics since June 2018. In addition, Dr. Stagliano has served as executive chair of Sundance Biosciences, Inc., a privately held biotechnology company, since January 2026. Dr. Stagliano previously served as Chief Executive Officer of True North until its acquisition by Bioverativ, Inc. in 2017. Prior to that, Dr. Stagliano served as Chief Executive Officer of iPierian, Inc., a privately held biotechnology company focused on neurodegenerative disease, from 2011 until its acquisition by Bristol-Myers Squibb Company (Bristol-Myers Squibb) in 2014. She also served as Chief Executive Officer of CytomX Therapeutics, Inc., a publicly held clinical-stage biopharmaceutical company, from 2008 to 2011. Dr. Stagliano earned her Ph.D. in neuroscience from the University of Miami School of Medicine and completed a postdoctoral fellowship at Harvard Medical School, as well as an M.S. in biomedical engineering and a B.S. in electrical engineering, both from Drexel University.
Our board of directors believes that Dr. Stagliano’s extensive leadership experience in the biotechnology field, her expertise in drug development, and her broad knowledge of the biopharmaceutical industry qualify her to serve on our board of directors.
Matthew Fust, M.B.A., has served as a member of our board of directors since November 2023. Mr. Fust has served on the board of directors of Atara Biotherapeutics, Inc. since March 2014, Crinetics Pharmaceuticals, Inc. since March 2018, Ultragenyx Pharmaceutical, Inc. since January 2014, and Neumora Therapeutics, Inc. since December 2020, each of which are publicly held biotechnology companies. Mr. Fust was previously Executive Vice President and Chief Financial Officer of Onyx Pharmaceuticals, Inc., a publicly held biopharmaceutical company, from January 2009 through its acquisition by Amgen Inc. (Amgen) in October 2013. Mr. Fust continued as an employee of Amgen until January 2014. From May 2003 to December 2008, Mr. Fust served as Chief Financial Officer at Jazz Pharmaceuticals, Inc., a publicly held specialty pharmaceutical company. Mr. Fust earned his M.B.A. from the Stanford Graduate School of Business and his B.A. in Accounting from the University of Minnesota Duluth.
Our board of directors believes that Mr. Fust’s extensive experience as a chief financial officer in the life sciences industry, his leadership and management experience, and his service as a director of other biopharmaceutical companies qualify him to serve on our board of directors.
154
Table of Contents
Thomas Geninatti, Ph.D., has served as a member of our board of directors since October 2025. Dr. Geninatti has served as a Principal of Nextech Invest AG, a biotechnology investment firm (Nextech Invest), since October 2025 and as a Senior Associate from September 2024 to October 2025. Dr. Geninatti has served as a member of the board of directors of Hexagon Bio, Inc., a privately held biotechnology company, since January 2025. Prior to Nextech Invest, he served in numerous roles of increasing responsibility at Gilead from December 2019 to September 2024, most recently serving as Senior Director of Corporate Development. Dr. Geninatti took a brief hiatus from Gilead from May 2022 to September 2022 to serve as Senior Director, Business Development & Licensing, Oncology at Bayer AG. Prior to joining Gilead in December 2019, Dr. Geninatti served as a senior associate project manager and consultant for Deallus Consulting Limited, a global life science consulting firm. Dr. Geninatti earned a Ph.D. in bioengineering from a joint program between the University of the Chinese Academy of Sciences and the Houston Methodist Research Institute, as well as B.S. and M.S. degrees in Biomedical Engineering from Politecnico di Torino, Italy.
Our board of directors believes that Dr. Geninatti’s experience in venture capital and the life science industry qualifies him to serve on our board of directors.
Carl L. Gordon, Ph.D., C.F.A., has served as a member of our board of directors since March 2019. Dr. Gordon is a Managing Partner at OrbiMed Advisors LLC, an investment firm, where he has served since January 1998. Dr. Gordon has served on the board of directors of Compass Therapeutics Inc., a publicly held biopharmaceutical company, since September 2015, BlossomHill Therapeutics, Inc., a publicly held biopharmaceutical company, since February 2021, Scribe Therapeutics Inc., a publicly held biopharmaceutical company, since March 2021, Lomond Therapeutics Holdings, Inc., a privately held biotechnology company, since August 2024, and currently serves on the boards of directors of several other privately held companies. Dr. Gordon previously served on the boards of directors of several publicly held companies, including Adicet Bio, Inc. from August 2015 to April 2025, ArriVent Biopharma, Inc. from December 2022 to June 2025, Gemini Therapeutics, Inc. (which merged with Disc Medicine, Inc.) from April 2016 until December 2022, Keros Therapeutics, Inc. from March 2020 to March 2026, Kinnate Biopharma Inc. from December 2019 to April 2024, MBX Biosciences, Inc. from July 2020 to June 2025, ORIC Pharmaceuticals, Inc. from November 2015 to November 2021, Terns Pharmaceuticals, Inc. from October 2018 to February 2025, and Theseus Pharmaceuticals, Inc. from June 2018 to February 2024. Dr. Gordon earned a B.A. in chemistry from Harvard College, a Ph.D. in molecular biology from the Massachusetts Institute of Technology and was a Fellow at The Rockefeller University.
Our board of directors believes that Dr. Gordon’s scientific expertise, extensive business experience, and experience in venture capital and the life science industry, including as a board member of various biotechnology and pharmaceutical companies, qualify him to serve on our board of directors.
Beth Seidenberg, M.D., has served as a member of our board of directors since March 2019. Dr. Seidenberg is the founding Managing Director of Westlake BioPartners, a life sciences venture capital firm, formed September 2018. Dr. Seidenberg is also a general partner at Kleiner Perkins Caufield and Byers, LLC, a venture capital firm, where she has primarily focused on life sciences investing since May 2005. From February 2002 to December 2004, Dr. Seidenberg served as the Senior Vice President, Head of Global Development and Chief Medical Officer at Amgen, a publicly held biotechnology company. Prior to that, Dr. Seidenberg was a senior executive in research and development at Bristol-Myers Squibb, a publicly held biopharmaceutical company, and at Merck & Co., Inc., a publicly held biopharmaceutical company. Dr. Seidenberg has served on the board of directors of Latigo Biotherapeutics, Inc., a publicly held biopharmaceutical company, since July 2024, Vera Therapeutics, Inc., a publicly held biotechnology company, since June 2016, Kyverna Therapeutics, Inc., a publicly held biopharmaceutical company, since September 2018, Sagimet Biosciences Inc., a publicly held biotechnology company, since April 2007, and several other privately held life sciences companies. Dr. Seidenberg previously served on the board of directors of Acelyrin, Inc., a publicly held biopharmaceutical company, from October 2020 to May 2025, Atara Biotherapeutics, Inc., a publicly held biopharmaceutical company, from August 2012 to June 2023, and Progyny, Inc., a publicly held fertility benefit management company, from May 2010 to November 2024. Dr. Seidenberg earned her B.A. in Biology and Anthropology from Barnard College and her M.D. from the University of Miami School of Medicine and completed her post-graduate training at The Johns Hopkins University, George Washington University, and the National Institutes of Health.
155
Table of Contents
Our board of directors believes that Dr. Seidenberg’s extensive experience as a life sciences investor, her significant senior industry leadership experience in clinical and drug development, and her broad service on the boards of biotechnology companies qualify her to serve on our board of directors.
Family Relationships and Other Arrangements
There are no family relationships among the directors and executive officers. Pursuant to our amended and restated voting agreement, which will terminate upon the closing of this offering, the following directors were designated as members of our board of directors:
| ∎ | Dr. Dong, designated pursuant to her service as our Chief Executive Officer; |
| ∎ | Dr. Geninatti, designated by Nextech VIII SCSp (Nextech); |
| ∎ | Dr. Seidenberg, designated by Westlake BioPartners Fund II, L.P. (Westlake); |
| ∎ | Dr. Gordon, designated by OrbiMed Private Investments VII, LP (OPI VII); |
| ∎ | Mr. Fust, designated by unanimous approval of the other members of our board of directors; and |
| ∎ | Dr. Stagliano, designated by unanimous approval of the other members of our board of directors. |
Composition of our Board of Directors
Our business and affairs are organized under the direction of our board of directors, which currently consists of six members. Drs. Geninatti and Seidenberg have notified us that they each intend to resign from our board of directors effective as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. Following Drs. Geninatti’s and Seidenberg’s resignations, our board of directors will consist of four members. Such resignations from our board of directors are an ordinary course transition in connection with our public company preparedness. There have been no disagreements between us and the resigning directors on any matter relating to our operations, policies or practices. The primary responsibilities of our board of directors are to provide oversight, strategic guidance, counseling, and direction to our management. Our board of directors meets on a regular basis and on an ad hoc basis as required. In accordance with the terms of our amended and restated certificate of incorporation and amended and restated bylaws, which will become effective upon the closing of this offering, we will divide our board of directors into three classes, as follows:
| ∎ | Class I, which will consist of Drs. Gordon and Stagliano, whose terms will expire at our first annual meeting of stockholders to be held following this offering; |
| ∎ | Class II, which will consist of Mr. Fust, whose term will expire at our second annual meeting of stockholders to be held following this offering; and |
| ∎ | Class III, which will consist of Dr. Dong, whose term will expire at our third annual meeting of stockholders to be held following this offering. |
At each annual meeting of stockholders to be held after the initial classification, the successors to directors whose terms then expire will serve until the third annual meeting following their election and until their successors are duly elected and qualified. The authorized size of our board of directors is currently seven members. The authorized number of directors may be changed only by resolution of our board of directors. Any additional directorships resulting from an increase in the number of directors will be distributed among three classes so that, as nearly as possible, each class will consist of one-third of the directors. This classification of our board of directors may have the effect of delaying or preventing changes in our control or management.
Subject to the rights of any series of preferred stock to elect directors, our directors may only be removed for cause, which removal may be effected by the affirmative vote of the holders of at least 66-2/3% of our voting stock.
Board Leadership Structure
Our board of directors is currently chaired by Dr. Stagliano, who has authority, among other things, to call and preside over board of directors meetings, to set meeting agendas and to determine materials to be distributed to the board of directors. Accordingly, the Chairperson has substantial ability to shape the work of the board of directors. In addition, we have a separate chair for each committee of our board of directors. The chair of each committee is expected to report annually to our board of directors on the activities of their committee in fulfilling their responsibilities as detailed in their respective charters or specify any shortcomings should that be the case. We believe that the separation of responsibilities as among our President and Chief Executive Officer, Chairperson, and our committee chairs provides a balanced approach to managing the board of directors and overseeing our company. Our board of directors has concluded that our current leadership structure is appropriate at this time. However, our
156
Table of Contents
board of directors will continue to periodically review our leadership structure and may make such changes in the future as it deems appropriate.
Role of the Board in Risk Oversight
The audit committee of our board of directors is primarily responsible for overseeing our risk management processes on behalf of our board of directors. Going forward, we expect that the audit committee will receive reports from management periodically regarding our assessment of risks. In addition, the audit committee reports regularly to our board of directors, which also considers our risk profile. The audit committee and our board of directors focus on the most significant risks we face and our general risk management strategies.
While our board of directors oversees our risk management, management is responsible for day-to-day risk management processes. Our board of directors expects management to consider risk and risk management in each business decision, to proactively develop and monitor risk management strategies and processes for day-to-day activities, and to effectively implement risk management strategies adopted by the audit committee and our board of directors. We believe this division of responsibilities is the most effective approach for addressing the risks we face and that our board of directors’ leadership structure, which also emphasizes the independence of our board of directors in its oversight of its business and affairs, supports this approach.
Committees of our Board of Directors
Our board of directors has established or will establish an audit committee, a compensation committee, and a nominating and corporate governance committee. Our board of directors may establish other committees to facilitate the management of our business. The composition and functions of each committee are described below. Members serve on these committees until their resignation or until otherwise determined by our board of directors. Our board of directors may establish other committees as it deems necessary or appropriate from time to time. Each committee has adopted or will adopt a written charter that satisfies the applicable rules and regulations of the Sarbanes-Oxley Act, the SEC, and the listing standards of the Nasdaq Stock Market LLC (Nasdaq), which we will post on our website, www.electra-therapeutics.com, upon the closing of this offering. We have included our website in this prospectus solely as a textual reference. The information contained in, or accessible through, our website is not incorporated by reference into this prospectus, and you should not consider any information contained in, or that can be accessed through, our website as part of this prospectus or in deciding whether to purchase our common stock.
Audit Committee
Upon the closing of this offering, our audit committee will consist of Mr. Fust, Dr. Gordon, and Dr. Stagliano. Mr. Fust will serve as the chair of our audit committee. Our board of directors has determined that each of the members of our audit committee satisfies Nasdaq and SEC independence requirements. The primary functions of this committee include, among other things:
| ∎ | evaluating the performance, qualifications, and independence of our independent auditor and determining whether to retain our existing independent auditor or engage a new independent auditor; |
| ∎ | determining and approving the engagement of our independent auditor to perform audit services and any permissible non-audit services; |
| ∎ | monitoring the rotation of partners of our independent auditor on our engagement team as required by law; |
| ∎ | prior to engagement of any independent auditor, and at least annually thereafter, reviewing relationships that may reasonably be thought to bear on their independence, and assessing and otherwise taking the appropriate action to oversee the independence of our independent auditor; |
| ∎ | reviewing our annual and quarterly financial statements and reports, including the disclosures contained under the caption “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” and discussing the statements and reports with our independent auditor and management; |
| ∎ | reviewing, with our independent auditor and management, major issues that arise regarding accounting principles and financial statement presentation, including any significant changes in selection or application of accounting principles, significant regulatory or accounting initiatives or developments, as well as off-balance sheet structures, that may have a material impact on our financial statements; |
| ∎ | reviewing, with our management, the scope, adequacy, and effectiveness of our internal control over financial reporting; |
157
Table of Contents
| ∎ | reviewing with management and our independent auditor any earnings announcements and other public announcements regarding material developments; |
| ∎ | establishing procedures for the receipt, retention, and treatment of complaints received by us regarding accounting, internal accounting controls, or auditing matters, as well as other matters; |
| ∎ | preparing the audit committee report required by the rules of the SEC to be included in our annual proxy statement; |
| ∎ | reviewing, approving, or ratifying and providing oversight of any related-person transactions in accordance with our related-person transaction policy; |
| ∎ | reviewing and monitoring compliance with programs and policies designed to ensure adherence to applicable laws and our code of business conduct and ethics; |
| ∎ | reviewing and assessing our risk management, risk assessment, and major risk exposures with respect to financial, accounting, operational, tax, privacy and cybersecurity, and information technology risks; |
| ∎ | reviewing on a periodic basis our investment policy and related-person transactions policy; and |
| ∎ | reviewing and evaluating, at least on an annual basis, the performance of the audit committee and the adequacy of the audit committee charter. |
Our board of directors has determined that Mr. Fust qualifies as an “audit committee financial expert” within the meaning of SEC regulations and meets the financial sophistication requirements of the Nasdaq Listing Standards. In making this determination, our board has considered prior experience, business acumen, and independence. Both our independent registered public accounting firm and management periodically meet privately with our audit committee.
We believe that the composition and functioning of our audit committee complies with all applicable requirements of the Sarbanes-Oxley Act and all applicable SEC rules and regulations. We intend to comply with future requirements to the extent they become applicable to us.
Compensation Committee
Our compensation committee consists of Mr. Fust and Dr. Gordon. Dr. Gordon will serve as the chair of our compensation committee. Our board of directors has determined that each of the members of our compensation committee is a non-employee director, as defined in Rule 16b-3 promulgated under the Securities Exchange Act of 1934, as amended (Exchange Act), and satisfies the independence requirements. The primary functions of this committee include, among other things:
| ∎ | overseeing our overall compensation practices and objectives, and assessing whether our compensation practices establish appropriate incentives in light of our specific business objectives; |
| ∎ | reviewing and approving or, if it deems it appropriate, making recommendations to the full board of directors regarding the compensation and other terms of employment of our executive officers; |
| ∎ | reviewing and approving (or if it deems it appropriate, making recommendations to the full board of directors regarding) corporate goals and objectives relevant to the compensation of our executive officers and assessing their performance against these goals and objectives; |
| ∎ | reviewing and approving (or if it deems it appropriate, making recommendations to the full board of directors regarding) the establishment or modification of our equity-based compensation plans and any other of our incentive compensation plans; |
| ∎ | evaluating risks associated with our compensation policies and practices and assessing whether risks arising from our compensation policies and practices for our employees are reasonably likely to have a material adverse effect on us; |
| ∎ | reviewing and discussing with management our policies and practices related to our management of human capital resources and corporate culture; |
| ∎ | modifying and overseeing the compensation clawback or similar policies; |
| ∎ | reviewing and making recommendations to the full board of directors regarding the form and amount of director compensation; |
158
Table of Contents
| ∎ | providing recommendations to the full board of directors on compensation-related proposals to be considered at our annual meeting of stockholders, including any applicable advisory votes on executive compensation and the frequency of such votes, incentive and other compensation plans, and amendments to such plans; |
| ∎ | reviewing and assessing the independence of compensation consultants, legal counsel, and other advisors as required by Section 10C of the Exchange Act; |
| ∎ | administering our incentive and equity-based compensation plans; |
| ∎ | establishing policies with respect to equity compensation arrangements; |
| ∎ | reviewing and approving the list of companies, if any, to be included in any compensation peer group used to determine pay levels based on criteria the compensation committee deems appropriate; |
| ∎ | reviewing and approving (or if it deems it appropriate, making recommendations to the full board of directors regarding) any employment or post-employment agreement or arrangement (including severance and change in control benefits) applicable to any of our executive officers; |
| ∎ | reviewing with management and approving our disclosures under the caption “Compensation Discussion and Analysis” to be included in our annual proxy statement or annual report on Form 10-K, in accordance with the rules of the SEC; |
| ∎ | preparing the compensation committee report on executive officer compensation as required by the SEC to be included in our annual proxy statement or annual report on Form 10-K filed with the SEC; |
| ∎ | reviewing with management and making recommendations to the full board of directors regarding the plans for succession of our Chief Executive Officer and other key executives; and |
| ∎ | reviewing and assessing on an annual basis the performance of the compensation committee and the compensation committee charter. |
We believe that the composition and functioning of our compensation committee complies with all applicable requirements of the Sarbanes-Oxley Act and all applicable SEC and Nasdaq rules and regulations. We intend to comply with future requirements to the extent they become applicable to us.
Nominating and Corporate Governance Committee
Upon the closing of this offering, our nominating and corporate governance committee will consist of Mr. Fust and Dr. Stagliano. Dr. Stagliano will serve as the chair of our nominating and corporate governance committee. Our board of directors has determined that each of the members of this committee satisfies the independence requirements. The primary functions of this committee include, among other things:
| ∎ | identifying, reviewing, screening, and evaluating candidates to serve on our board of directors consistent with criteria approved by our board of directors; |
| ∎ | determining the minimum qualifications for service on our board of directors; |
| ∎ | overseeing evaluations of director performance on the board and applicable committees of the board and determining whether continued service on our board is appropriate; |
| ∎ | reviewing and making recommendations to the board of directors concerning the size, structure, composition, and functioning of the board and its committees; |
| ∎ | reviewing any stockholder proposals submitted for inclusion in our proxy statement and recommending to the full board of directors any statements by us in response, and considering stockholder nominees for election to our board of directors at our annual meeting of stockholders; |
| ∎ | considering and assessing the independence of members of our board of directors and executive officers; |
| ∎ | developing and making recommendations to the Board regarding corporate governance guidelines, periodically reviewing these guidelines, and recommending to our board of directors any changes to such guidelines; |
| ∎ | overseeing our environmental, social, and governance strategies, targets, policies, performance, and reporting; and |
| ∎ | reviewing and assessing on an annual basis the performance of the nominating and corporate governance committee and the nominating and corporate governance committee charter. |
159
Table of Contents
We believe that the composition and functioning of our nominating and corporate governance committee complies with all applicable requirements of the Sarbanes-Oxley Act and all applicable SEC and Nasdaq rules and regulations. We intend to comply with future requirements to the extent they become applicable to us.
Compensation Committee Interlocks and Insider Participation
None of our current or former executive officers serve as a member of the compensation committee. Except for Dr. Dong, who has served as a member of the board of directors of Neuron23 since June 2022 where Dr. Stagliano currently serves as chief executive officer and chair of the board, none of our officers serve, or have served during the last completed fiscal year, on the board of directors or compensation committee, or other committee serving an equivalent function, of any other entity that has one or more of its executive officers serving as a member of our board of directors or our compensation committee.
Code of Business Conduct and Ethics
In connection with this offering, we have adopted a written code of business conduct and ethics that applies to our directors, officers, and employees, including our principal executive officer, principal financial officer, principal accounting officer or controller, or person performing similar functions. Following this offering, a current copy of the code will be available on the Corporate Governance section of our website, www.electra-therapeutics.com. We have included our website in this prospectus solely as a textual reference. The information contained in, or accessible through, our website is not incorporated by reference into this prospectus, and you should not consider any information contained in, or that can be accessed through, our website as part of this prospectus or in deciding whether to purchase our common stock.
Director Independence
Our board of directors has undertaken a review of the independence of each director. Based on information provided by each director concerning his or her background, employment, and affiliations, our board of directors has determined that with the exception of Dr. Dong, none of our directors have relationships that would interfere with the exercise of independent judgment in carrying out the responsibilities of a director and that each of these directors is “independent” as that term is defined under the applicable listing standards. In making these determinations, our board of directors considered the current and prior relationships that each non-employee director has with our company and all other facts and circumstances our board of directors deemed relevant in determining their independence, including the beneficial ownership of our shares held by each non-employee director and the transactions described in the section titled “Certain Relationships and Related Person Transactions.”
160
Table of Contents
EXECUTIVE AND DIRECTOR COMPENSATION
Our named executive officers for the year ended December 31, 2025, consisting of our principal executive officer and the next two most highly compensated executive officers, were:
| ∎ | Quehuong (Kathy) Dong, Pharm.D., M.B.A., our President and Chief Executive Officer; |
| ∎ | Kim-Hien Dao, D.O., Ph.D., our Chief Medical Officer; and |
| ∎ | Gary S. Koe, Ph.D., our Chief Technical Officer. |
Summary Compensation Table for the Year Ended December 31, 2025
The following table presents all of the compensation awarded to, earned by or paid to our named executive officers during the year ended December 31, 2025.
| NAME AND PRINCIPAL POSITION |
YEAR | SALARY ($) |
OPTION AWARDS ($) (1) |
NON-EQUITY INCENTIVE PLAN COMPENSATION ($) (2) |
TOTAL ($) |
|||||||||||||||
| Quehuong (Kathy) Dong, Pharm.D., M.B.A., President and Chief Executive Officer |
2025 | 485,000 | 283,037 | 250,000 | 1,018,037 | |||||||||||||||
| Kim-Hien Dao, D.O., Ph.D., Chief Medical Officer |
2025 | 400,107 | — | 147,040 | 547,147 | |||||||||||||||
| Gary S. Koe, Ph.D., Chief Technical Officer |
2025 | 389,870 | 188,731 | 133,264 | 711,865 | |||||||||||||||
| (1) | In accordance with SEC rules, this column reflects the aggregate grant date fair value of the stock options granted during the year ended December 31, 2025. These amounts have been computed in accordance with Financial Accounting Standards Board (FASB), Accounting Standards Codification (ASC) Topic 718. As required by SEC rules, the amounts shown exclude the impact of estimated forfeitures related to service-based vesting conditions. Assumptions used in the calculation of these amounts are described in Note 2 to our financial statements included elsewhere in this prospectus. These amounts do not reflect the actual economic value that may be realized by our named executive officers upon vesting or exercise of the stock options or the sale of the common stock underlying such awards. |
| (2) | The amounts disclosed represent performance-based bonuses earned with respect to the year ended December 31, 2025, as described in the subsection below titled “—Annual Bonuses.” |
Narrative to Summary Compensation Table
Annual Base Salary
Our named executive officers receive base salaries to compensate them for services rendered to us. The base salary payable to each named executive officer is intended to provide a fixed component of compensation, taking into account such named executive officer’s qualifications, experience, the scope of such named executive officer’s responsibilities and competitive market compensation paid by other companies for similar positions within the industry and geography. Base salaries are reviewed periodically and adjusted from time to time to realign salaries with market levels after taking into account individual responsibilities, performance and experience. The 2025 annual base salary rates for our named executive officers are set forth in the table below, and the base salaries ultimately earned by our named executive officers for 2025 are reported above in the “Salary” column of the Summary Compensation Table.
| NAME |
2025 BASE SALARY |
|||
| Quehuong (Kathy) Dong, Pharm.D., M.B.A. |
$ | 500,000 | (1) | |
| Kim-Hien Dao, D.O., Ph.D. |
$ | 400,107 | ||
| Gary S. Koe, Ph.D. |
$ | 395,575 | (2) | |
| (1) | In May 2025, our board of directors approved an increase to Dr. Dong’s annual base salary from $460,000 to $500,000 effective May 16, 2025. |
| (2) | In May 2025, our board of directors approved an increase to Dr. Koe’s annual base salary from $380,361 to $395,575 effective May 16, 2025 in connection with his promotion from SVP, Head of Tech Ops to Chief Technical Officer. |
161
Table of Contents
Annual Bonuses
In addition to base salaries, our named executive officers are eligible to receive annual bonuses based on the achievement of corporate and individual performance goals. For 2025, the corporate performance goals approved by our board of directors, included those related to research and development and clinical milestones and other corporate goals related to company finances and business development. Dr. Dong is eligible to receive an annual performance bonus weighted entirely on the achievement of corporate performance goals as determined by our board of directors. Drs. Dao and Koe are also eligible to receive annual bonuses based on the achievement of corporate and individual performance goals as determined by our board of directors, with 90% of the bonus based on achievement of corporate goals and 10% based on individual performance. These annual bonuses are paid out of a maximum bonus pool established by our board of directors (the bonus pool). For the year ended December 31, 2025, annual cash bonus targets, as a percentage of base salary, were 50% for Dr. Dong, 35% for Dr. Dao, and, with respect to Dr. Koe, 30% of his base salary through May 15, 2025 for the period he served as SVP, Head of Tech Ops and 35% for remainder of the year following his promotion to Chief Technical Officer. Following its assessment of 2025 corporate performance and, as applicable, individual performance, the board of directors determined that Dr. Dong had earned a 2025 annual bonus at 100% of target, Dr. Dao had earned a 2025 annual bonus at 105% of target, and Dr. Koe had earned a 2025 annual bonus on a blended basis at 103% of target. The annual bonuses ultimately earned by our named executive officers for 2025 are reported above in the “Non-Equity Incentive Plan Compensation” column of the Summary Compensation Table.
Equity-based Incentive Awards
Our equity-based incentive awards are designed to align the interests of our stockholders with those of our employees, including our named executive officers. The board of directors or an authorized committee thereof is responsible for approving equity grants.
Prior to this offering, we have granted stock options pursuant to our 2022 Stock Plan (2022 Plan) to certain of our executives, including our named executive officers. In addition, pursuant to the Electra Therapeutics LLC 2023 Equity Incentive Plan (2023 Plan), we granted, through Electra Therapeutics LLC (Electra LLC), Drs. Dong and Koe common units of Electra LLC intended to qualify as “profits interests” for U.S. federal income tax purposes. In connection with our spin-out from Star Therapeutics LLC, Electra LLC was formed as a holding company whose sole asset is the shares of our Series A redeemable convertible preferred stock. Service providers to us or Electra LLC were eligible to receive Profits Interest Unit (as defined below) awards under the 2023 Plan. Prior to the closing of this offering, Electra LLC expects to distribute shares of our Series A redeemable convertible preferred stock to the unit holders of Electra LLC, including to holders of outstanding profits interest units of Electra LLC (the LLC Distribution), which includes Dr. Dong and Dr. Koe. In connection with the offering, the Series A redeemable convertible preferred stock will convert into shares of common stock. Dr. Koe’s shares will be fully vested on issuance. In connection with the LLC Distribution, Dr. Dong and the Company will enter into a Stock Restriction Agreement such that 72,949 shares of Series A redeemable convertible preferred stock she receives in the LLC Distribution (Dr. Dong’s Unvested Shares), which reflects the number of shares of Series A redeemable convertible preferred stock that will be distributed to Dr. Dong with respect to her unvested Profits Interest Units of Electra LLC (assuming that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock), will be subject to vesting on substantially the same vesting schedule that applied to her Profits Interest Units, which means that 1/13th of Dr. Dong’s Unvested Shares will vest each month commencing from September 16, 2026, subject to her continuous service with us through each such vesting date. Additionally, in connection with the LLC Distribution, Dr. Dong will receive 381,419 shares of Series A redeemable convertible preferred stock (assuming that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock), which will be distributed to Dr. Dong with respect to her vested Profits Interest Units, will be fully vested, and will not be subject to the Stock Restriction Agreement.
Following this offering, we will grant equity awards under the terms of our 2026 Equity Incentive Plan (2026 Plan, and together with the 2022 Stock Plan, the equity plans). The terms of our equity plans and the 2023 Plan are described below under the subsection titled “—Equity Benefit Plans.” Our named executive officers received the following stock option awards under our 2022 Plan in the year ended December 31, 2025:
162
Table of Contents
On November 13, 2025, our board of directors granted option awards to each of Dr. Dong and Dr. Koe, covering 70,903 shares of our common stock and 47,269 shares of our common stock, respectively. These options are early exercisable, and have an exercise price of $4.96 per share and vest over a period of four years, with 25% of the option vesting on the first anniversary of the vesting commencement date and the remaining 75% of the option vesting ratably on a monthly basis thereafter, subject to Drs. Dong’s and Koe’s, as applicable, continued service with us through each vesting date.
Additionally, on February 13, 2026, our board of directors granted an option award to each of Dr. Dong, Dr. Dao and Dr. Koe, covering 759,198, 309,676 and 231,941, shares of our common stock, respectively. Each award has an exercise price of $4.96 per share and vests ratably on a monthly basis over four years, subject to the respective named executive officer’s continued service with us through each vesting date. Each option is also subject to early exercise and is fully exercisable as of the grant date.
On February 13, 2026, our board of directors also granted an option award to Dr. Dong covering 228,001 shares of our common stock. The award has an exercise price of $4.96 per share and vests ratably on a monthly basis over four years, subject to and contingent on (i) Dr. Dong’s continued service with us through each vesting date and (ii) the consummation of a change in control (as defined in the Dong Agreement, as defined below) on or before the later of (a) September 30, 2027 and (b) the date that is three months following the date our board of directors is presented with the ipsoprubart sHLH pivotal topline comparative data. The option is also subject to early exercise and is fully exercisable as of the grant date. On September 11, 2026, we entered into an Option Cancellation Agreement with Dr. Dong that will be effective concurrently with and contingent upon the granting of the IPO Option Grants, pursuant to which the Company and Dr. Dong mutually agreed to cancel this option in exchange for receiving the options awarded to her in the IPO Option Grants. The board determined that the change in control vesting component was no longer relevant in light of the offering.
Stock options granted to our named executive officers are subject to potential acceleration of vesting in connection with a change of control, as described below under the subsection titled “—Potential Payments Upon Termination or Change of Control.”
Outstanding Equity Awards as of December 31, 2025
The following table presents the outstanding equity awards held by each named executive officer as of December 31, 2025.
| OPTION AWARDS (1) | STOCK AWARDS (2) | |||||||||||||||||||||||||||||||
| NAME |
GRANT DATE | VESTING COMMENCEMENT DATE |
NUMBER OF SECURITIES UNDERLYING UNEXERCISED OPTIONS EXERCISABLE (#) |
NUMBER OF SECURITIES UNDERLYING UNEXERCISED OPTIONS UNEXERCISABLE (#) |
OPTION EXERCISE PRICE ($) |
OPTION EXPIRATION DATE |
NUMBER OF SHARES OR UNITS OF STOCK THAT HAVE NOT VESTED (#) |
MARKET VALUE OF SHARES OF UNITS OR STOCK THAT HAVE NOT VESTED ($) (3) |
||||||||||||||||||||||||
| Quehuong (Kathy) Dong, Pharm.D., M.B.A. |
03/24/2022 | 03/24/2022 | (4) | 59,086 | — | 5.23 | 03/23/2032 | — | — | |||||||||||||||||||||||
| 11/17/2023 | 10/16/2023 | (4) | 281,987 | — | 4.40 | 11/16/2033 | — | — | ||||||||||||||||||||||||
| 11/17/2023 | 10/16/2023 | (5) | — | — | — | — | 148,487 | 112,850 | ||||||||||||||||||||||||
| 02/02/2024 | 10/16/2023 | (5) | — | — | — | — | 555,335 | 422,055 | ||||||||||||||||||||||||
| 11/13/2025 | 05/08/2025 | (4) | 70,903 | — | 4.96 | 11/12/2035 | — | — | ||||||||||||||||||||||||
| Kim-Hien Dao, D.O., Ph.D. |
08/09/2023 | 04/03/2023 | (4) | 62,631 | — | 4.40 | 08/08/2033 | — | — | |||||||||||||||||||||||
| 11/17/2023 | 01/01/2024 | (4) | 23,634 | — | 4.40 | 11/16/2033 | — | — | ||||||||||||||||||||||||
| 05/09/2024 | 05/16/2024 | (4) | 103,992 | — | 5.05 | 05/08/2034 | — | — | ||||||||||||||||||||||||
| 11/14/2024 | 11/14/2024 | (4) | 47,269 | — | 5.05 | 11/13/2034 | — | — | ||||||||||||||||||||||||
| Gary S. Koe, Ph.D. |
03/24/2022 | 03/24/2022 | (4) |
|
35,451 |
|
— | 5.23 | 03/23/2032 | — | — | |||||||||||||||||||||
| 11/17/2023 | 01/01/2024 | (4) | 47,269 | — | 4.40 | 11/16/2033 | — | — | ||||||||||||||||||||||||
| 05/09/2024 | 05/16/2024 | (4) | 18,907 | — | 5.05 | 05/08/2034 | — | — | ||||||||||||||||||||||||
| 11/13/2025 | 05/16/2025 | (4) | 47,269 | — | 4.96 | 11/12/2035 | — | — | ||||||||||||||||||||||||
| (1) | All of the option awards were granted under the 2022 Plan, the terms of which plan is described below under “—Equity Benefit Plans—2022 Stock Plan.” |
163
Table of Contents
| (2) | Represents Profits Interest Units. All of the Profits Interest Units were granted under the 2023 Plan, the terms of which are described below under the subsection titled “—Equity Benefit Plans—Electra Therapeutics LLC 2023 Equity Incentive Plan.” Each of the Profits Interest Units has a distribution threshold (i.e., profits interest threshold amount), which represents the fair market value of an Electra LLC common unit at the time of grant. Holders of Profits Interest Units only participate in distributions from Electra LLC to the extent the value of the distributions made on all other units of Electra LLC exceed the applicable profits interest threshold amount. Prior to the closing of this offering, Electra LLC expects to complete the LLC Distribution, and any shares of Series A redeemable convertible preferred stock received in the LLC Distribution will automatically convert into shares of our common stock immediately prior to the closing of this offering. Dr. Dong holds an aggregate of 2,345,871 Profits Interest Units (inclusive of the 703,822 Profits Interest Units listed in the table above), of which (i) 252,910 Profits Interest Units have a distribution threshold of $0.46 per unit, all of which are fully vested and not reflected in the above table, (ii) 557,351 Profits Interest Units have a distribution threshold of $1.21 per unit, all of which are fully vested and not reflected in the table above, and (iii) 1,535,610 Profits Interest Units have a distribution threshold of $1.77 per unit, 831,788 of which were vested as of December 31, 2025 and not reflected in the table above. Dr. Dong’s Unvested Shares will be subject to vesting as described above, and all shares received with respect to her vested Profits Interest Units will be fully vested. Dr. Koe holds an aggregate of 270,087 Profits Interest Units, of which (x) 126,455 Profits Interest Units have a distribution threshold of $0.49 per unit, all of which are fully vested and not reflected in the table above, and (y) 143,632 Profits Interest Units have a distribution threshold of $1.21 per unit, all of which are fully vested and not reflected in the table above. |
| (3) | The market value is based on the fair market value of our Profits Interest Units as of December 31, 2025 of $2.53 per unit less the aggregate distribution threshold of the unvested Profits Interest Units. |
| (4) | 25% of the total shares vest on the first anniversary of the vesting commencement date and the remaining 75% of the total shares vest ratably in monthly installments over the three-year period thereafter, subject to continuous service with us through each vesting date. In addition, the shares subject to this option are subject to acceleration as set forth in our Severance Benefit Plan (as described below). For more information, please see the subsection titled “—Potential Payments Upon Termination or Change of Control” below. Each option is also subject to early exercise and is fully exercisable as of the grant date. |
| (5) | 25% of the total units vest on the first anniversary of the vesting commencement date and the remaining 75% of the total units vest ratably in monthly installments over the three-year period thereafter, subject to continuous service with us through each vesting date. In addition, if (a) Electra LLC is subject to a change in control before Dr. Dong’s service with Electra LLC terminates and (b) Dr. Dong is subject to an involuntary termination within 12 months after that change in control, then 100% of the units shall vest. |
IPO Grants
At the time of the effectiveness of the 2026 Plan, our board of directors will grant options to purchase shares of our common stock to our named executive officers, with an exercise price equal to the initial public offering price per share. As of such date, Drs. Dong, Koe and Dao will be granted 927,155, 155,979, and 155,979 options to purchase shares of our common stock, respectively. The shares underlying the options vest ratably in monthly installments over four years, subject to continuous service with us through each vesting date.
In addition, to allow Drs. Dong and Koe to retain a substantially similar ownership and economic position that applied to their respective Profit Interest Units prior to the LLC Distribution, at the time of the effectiveness of the 2026 Plan, our board of directors has determined to grant Dr. Dong an option to purchase 199,890 shares of our common stock and Dr. Koe an option to purchase 11,800 shares of our common stock, in each case, with an exercise price equal to the per share price as set forth on the cover of the prospectus to this offering. The vesting of these options will have substantially the same vesting terms as applied to the Profits Interests Units to which they correspond. As such, with respect to Dr. Dong, 167,798 shares underlying Dr. Dong’s option will be vested in full as of the date of grant, and the remaining 32,092 shares will vest as to 1/13th of the shares monthly commencing from September 16, 2026, subject to her continuous service with us through each such vesting date, and with respect to Dr. Koe, all of the shares underlying Dr. Koe’s option will be fully vested as of the date of grant.
Employment Arrangements with Our Named Executive Officers
We have entered into an employment offer letter with each of our named executive officers. The material terms of each such agreement are described below. The employment of each of our named executive officers is “at will” and may be terminated at any time.
Quehuong (Kathy) Dong, Pharm.D., M.B.A.
Effective September 2023, we entered into an offer letter, as amended in November 2024 and May 2026, with Dr. Dong in connection with her service as our President and Chief Executive Officer (Dong Agreement). Under the Dong Agreement, Dr. Dong’s initial annual base salary was set at $425,000 (which has subsequently been increased to $517,500 effective January 1, 2026), subject to review and adjustment from time to time. The Dong Agreement also provides that Dr. Dong is eligible to receive an annual discretionary bonus at a target amount of 40% of her then-current base salary (which was subsequently increased to 50% of her then-current base salary effective beginning with her 2025 bonus opportunity), based on achievement of criteria determined by our board of directors.
164
Table of Contents
On August 20, 2026, we entered into a confirmatory offer letter with Dr. Dong in connection with her service as our President and Chief Executive Officer (A&R Dong Agreement), which will be effective immediately prior to and contingent upon the effectiveness of the registration statement of which this prospectus forms a part. Under the A&R Dong Agreement, Dr. Dong’s annual base salary was set at $625,000. The A&R Dong Agreement also provides that Dr. Dong is eligible to receive an annual discretionary bonus at a target amount of 55% of her then-current base salary, based upon our and Dr. Dong’s achievement of objectives and milestones to be approved by our board of directors.
In the event of certain qualifying terminations of her employment, Dr. Dong will be entitled to receive severance benefits, as described below under “—Potential Payments Upon Termination or Change of Control.”
Gary S. Koe, Ph.D.
Effective November 2023, we entered into an offer letter, as amended in May 2025, with Dr. Koe in connection with his service as Chief Technical Officer (Koe Agreement). Under the Koe Agreement, Dr. Koe’s initial annual base salary was $395,575 (which has subsequently been increased to $419,310, effective January 1, 2026). The Koe Agreement also provides that Dr. Koe is eligible to receive an annual discretionary bonus at a target amount of 35% of his then-current base salary (which was subsequently increased to 40% of his then-current base salary effective beginning with his 2026 bonus opportunity), based on criteria to be established by our Chief Executive Officer and approved by our board of directors. The Koe Agreement also provides for an option to purchase 47,269 shares of our common stock, as described further under “—Equity-based Incentive Awards.”
On August 20, 2026, we entered into a confirmatory offer letter with Dr. Koe in connection with his service as our Chief Technical Officer (A&R Koe Agreement), which will be effective immediately prior to and contingent upon the effectiveness of the registration statement of which this prospectus forms a part. Under the A&R Koe Agreement, Dr. Koe’s annual base salary was set at $450,000, subject to review and adjustment from time to time. The A&R Koe Agreement also provides that Dr. Koe is eligible to receive an annual discretionary bonus at a target amount of 40% of his then-current base salary, based upon our and Dr. Koe’s achievement of objectives and milestones to be approved by our board of directors.
In the event of certain qualifying terminations of his employment, Dr. Koe will be entitled to receive severance benefits, as described below under “—Potential Payments Upon Termination or Change of Control.”
Kim-Hien Dao, D.O., Ph.D.
Effective November 2023, we entered into an offer letter, as amended in November 2024 and May 2026, with Dr. Dao in connection with her service as our Vice President, Clinical Development, which continues to govern her employment as our Chief Medical Officer (Dao Agreement). Under the Dao Agreement, Dr. Dao’s initial annual base salary was set at $340,000 (which has subsequently been increased to $432,116 effective January 1, 2026), subject to review and adjustment from time to time. The Dao Agreement also provides that Dr. Dao is eligible to receive an annual discretionary bonus at a target amount of 30% of her then-current base salary (which was subsequently increased to 40% of her then-current base salary effective beginning with her 2026 bonus opportunity), based on criteria to be established by our Chief Executive Officer and approved by our board of directors.
On August 20, 2026, we entered into a confirmatory offer letter with Dr. Dao in connection with her service as our Chief Medical Officer (A&R Dao Agreement), which will be effective immediately prior to and contingent upon the effectiveness of the registration statement of which this prospectus forms a part. Under the A&R Dao Agreement, Dr. Dao’s annual base salary was set at $500,000, subject to review and adjustment from time to time. The A&R Dao Agreement also provides that Dr. Dao is eligible to receive an annual discretionary bonus at a target amount of 40% of her then-current base salary, based upon our and Dr. Dao’s achievement of objectives and milestones to be approved by our board of directors.
In the event of certain qualifying terminations of her employment, Dr. Dao will be entitled to receive severance benefits, as described below under the subsection titled “—Potential Payments Upon Termination or Change of Control.”
165
Table of Contents
Potential Payments Upon Termination or Change of Control
Regardless of the manner in which a named executive officer’s service terminates, each named executive officer is entitled to receive amounts earned during the term of service, including unpaid base salary accrued through the termination date. In addition, as of December 31, 2025 each of our named executive officers was entitled to certain severance benefits under their respective employment agreements, as described below, subject to their execution of a general release of claims in the form prescribed by us. We have adopted a severance and change in control plan to be effective upon the closing of this offering which will supersede any severance benefits or entitlements provided in the existing employment agreements.
The Dong Agreement provides that, if Dr. Dong’s employment is terminated by us without “cause” or Dr. Dong resigns for “good reason,” she will be entitled to receive continued payment of her then-current base salary for nine (9) months and payment of continued group healthcare benefit premiums under COBRA for up to nine (9) months. In addition, if we consummate a change in control and Dr. Dong is subject to an involuntary termination within 12 months following such change in control, then 100% of the then-unvested shares subject to her outstanding options under the 2022 Plan will immediately vest and become exercisable, and 100% of the then-unvested Common Units of Electra LLC held by Dr. Dong will immediately vest.
The Koe Agreement provides that, if Dr. Koe’s employment is terminated by us without “cause” or Dr. Koe resigns for “good reason,” he will be entitled to receive continued payment of his then-current base salary for six (6) months and reimbursement of continued group healthcare benefit premiums under COBRA for up to six (6) months. In addition, if we consummate a change in control and Dr. Koe is subject to an involuntary termination within 12 months following such change in control, then the vesting and exercisability of all outstanding time-based stock options and other time-based equity awards covering our common stock held by Dr. Koe will accelerate in full, conditioned upon the actual consummation of such change in control.
The Dao Agreement provides that, if Dr. Dao’s employment is terminated by us without “cause” or Dr. Dao resigns for “good reason,” she will be entitled to receive continued payment of her then-current base salary for six (6) months and payment of continued group healthcare benefit premiums under COBRA for up to six (6) months. In addition, if we consummate a change in control and Dr. Dao is subject to an involuntary termination within 12 months following such change in control, then 100% of the then-unvested shares subject to her outstanding options under the 2022 Plan will immediately vest.
Severance Plan
Upon and following this offering, each of our named executive officers will be eligible to receive severance benefits upon certain terminations of employment under the terms of our Severance Plan, which was adopted by our board of directors in August 2026 (Severance Plan) and which will be effective upon the execution and delivery of the underwriting agreement related to this offering. The Severance Plan supersedes and replaces in its entirety any prior severance benefits included in the Dong Agreement, the Koe Agreement, and the Dao Agreement.
Upon a termination without Cause or resignation for Good Reason (each as defined in our Severance Plan) outside of the period beginning 3 months prior to and ending 12 months following the effective date of a change in control (CIC Period), each of our named executive officers will be entitled to receive the following severance benefits: (i) cash severance in the form of salary continuation over the specified severance period (12 months for Dr. Dong and 9 months for Drs. Koe and Dao) and (ii) payment of COBRA premiums for the duration of the applicable severance period (12 months for Dr. Dong and 9 months for Drs. Koe and Dao).
Upon a termination without Cause or resignation for Good Reason during the applicable CIC Period, each of our named executive officers will be entitled to receive the following severance benefits in lieu of those described above: (i) a lump sum cash payment equal to a specified portion of the named executive officer’s base salary (18 months for Dr. Dong and 12 months for Drs. Koe and Dao), (ii) a lump sum cash payment equal to a specified percentage of the named executive officer’s annual target bonus for the year in which the change in control termination occurs (150% for Dr. Dong and 100% for Drs. Koe and Dao), (iii) payment of COBRA premiums for the duration of the applicable severance period (18 months for Dr. Dong and 12 months for Drs. Koe and Dong), and (iv) accelerated vesting of all then-outstanding and unvested time-based equity awards held by the named executive officer.
166
Table of Contents
All severance benefits under our Severance Plan are subject to the named executive officer’s execution (and non-revocation) of an effective release of claims against us. If any payments or benefits provided under our Severance Plan or otherwise would constitute “parachute payments” within the meaning of Section 280G of the Code and would be subject to the excise tax imposed by Section 4999 of the Code, then our named executive officers will receive the greater of (x) the largest portion of such payments that would not be subject to the excise tax, or (y) the largest portion, up to and including the total, of the payments, in each case, whichever produces the greater after-tax benefit to the executive. Our Severance Plan does not provide any excise tax gross-up payments.
Other Compensation and Benefits
All of our current named executive officers are eligible to participate in our employee benefit plans, which includes medical, dental, vision, and life insurance coverage, in each case on the same basis as all of our other employees. We pay a portion of the premiums for the medical, dental, and vision insurance, including for our named executive officers. We generally do not provide perquisites or personal benefits to our named executive officers. In addition, we provide the opportunity to participate in a 401(k) plan to our employees, including each of our named executive officers, as discussed in the subsection titled “—401(k) Plan” below.
401(k) Plan
Our named executive officers are eligible to participate in a defined contribution retirement plan that provides eligible employees with an opportunity to save for retirement on a tax advantaged basis. Eligible employees may defer eligible compensation on a pre-tax or after-tax (Roth) basis, up to the statutorily prescribed annual limits on contributions under the Internal Revenue Code of 1986, as amended (Code). Contributions are allocated to each participant’s individual account and are then invested in selected investment alternatives according to the participants’ directions. The 401(k) plan provides for discretionary matching contributions; we do not currently provide a match under the 401(k) plan.
Clawback Policy
In connection with this offering, we have adopted a compensation recovery policy that is compliant with the Nasdaq Listing Rules, as required by the Dodd-Frank Act, to be effective upon the consummation of this offering.
Equity Benefit Plans
We believe that our ability to grant equity-based awards is a valuable and necessary compensation tool that aligns the long-term financial interests of our employees, consultants and directors with the financial interests of our stockholders. In addition, we believe that our ability to grant stock options and other equity-based awards helps us to attract, retain and motivate employees, consultants and directors, and encourages them to devote their best efforts to our business and financial success. The principal features of our equity incentive plans are summarized below. These summaries are qualified in their entirety by reference to the actual text of the plans, which are filed as exhibits to the registration statement of which this prospectus forms a part.
2026 Equity Incentive Plan
Our board of directors adopted our 2026 Plan in August 2026 and our stockholders approved our 2026 Plan in September 2026. Our 2026 Plan is a successor to our 2022 Plan and will become effective on the execution of the underwriting agreement related to this offering. Once our 2026 Plan becomes effective, no further grants will be made under our 2022 Plan.
Types of Awards. Our 2026 Plan provides for the grant of incentive stock options (ISOs) to employees, including employees of any parent or subsidiary, and for the grant of nonstatutory stock options (NSOs), stock appreciation rights, restricted stock awards, restricted stock unit awards, performance awards and other forms of stock awards to employees, directors, and consultants, including employees and consultants of our affiliates.
Authorized Shares. Initially, the maximum number of shares of our common stock that may be issued under our 2026 Plan after it becomes effective will be 13,446,281 shares, which is the sum of: (i) 7,697,039 new shares, plus (ii) up to 5,749,242 shares subject to outstanding awards under the 2022 Plan that, following the effective date of the 2026 Plan, are not issued because such awards expire, are settled in cash, or are forfeited, or such shares are withheld to
167
Table of Contents
satisfy the exercise price thereof or to satisfy tax withholding obligations. In addition, the number of shares of our common stock reserved for issuance under our 2026 Plan will automatically increase on January 1 of each calendar year, starting on January 1, 2027 (assuming the 2026 Plan becomes effective in 2026) through January 1, 2036, in an amount equal to 5.0% of the total number of shares of our common stock (including shares issuable upon exercise, conversion, or exchange of all then-outstanding prefunded warrants, or other rights exercisable for or convertible into, directly or indirectly, shares of common stock (whether vested or unvested), but excluding shares issuable upon the exercise of then-outstanding options and the shares reserved and available for issuance under our then-existing equity incentive plan(s) and employee stock purchase plan(s), if any) outstanding on December 31 of the preceding calendar year, or a lesser number of shares determined by our board of directors. The maximum number of shares of our common stock that may be issued on the exercise of ISOs under our 2026 Plan is 40,338,843 shares.
Shares subject to stock awards granted under our 2026 Plan that expire or terminate without being exercised in full, or that are paid out in cash rather than in shares, do not reduce the number of shares available for issuance under our 2026 Plan. Additionally, shares become available for future grant under our 2026 Plan if they were issued under stock awards under our 2026 Plan and we repurchase them or they are forfeited. This includes shares used to pay the exercise price of a stock award or to satisfy the tax withholding obligations related to a stock award.
Plan Administration. Our board of directors, or a duly authorized committee of our board of directors (referred to herein as the plan administrator), will administer our 2026 Plan. Our board of directors may also delegate to one or more persons or bodies the authority to do one or more of the following (i) designate recipients (other than officers) of specified stock awards provided that no person or body may be delegated authority to grant a stock award to themselves; (ii) determine the number of shares subject to such stock award; and (iii) determine the terms of such stock awards. Under our 2026 Plan, our board of directors has the authority to determine and amend the terms of awards and underlying agreements, including recipients, the exercise, purchase or strike price of stock awards, if any, the number of shares subject to each stock award, the vesting schedule applicable to the awards, together with any vesting acceleration and the form of consideration, if any, payable on exercise or settlement of the award.
In addition, subject to the terms of the 2026 Plan, the administrator also has the power to modify outstanding awards under our 2026 Plan, including the authority to reprice any outstanding option or stock appreciation right, cancel and re-grant any outstanding option or stock appreciation right in exchange for new stock awards, cash or other consideration, or take any other action that is treated as a repricing under generally accepted accounting principles, with the consent of any materially adversely affected participant.
Stock Options. ISOs and NSOs are granted under stock option agreements adopted by the plan administrator. The plan administrator determines the exercise price for stock options, within the terms and conditions of the 2026 Plan, provided that the exercise price of a stock option generally cannot be less than 100% of the fair market value of our common stock on the date of grant. Options granted under the 2026 Plan vest at the rate specified in the stock option agreement as determined by the plan administrator.
Tax Limitations on ISOs. The aggregate fair market value, determined at the time of grant, of our common stock with respect to ISOs that are exercisable for the first time by an option holder during any calendar year under all of our stock plans may not exceed $100,000. Options or portions thereof that exceed such limit will generally be treated as NSOs. No ISO may be granted to any person who, at the time of the grant, owns or is deemed to own stock possessing more than 10% of our total combined voting power or that of any of our affiliates unless (i) the option exercise price is at least 110% of the fair market value of the stock subject to the option on the date of grant; and (ii) the option is not exercisable after the expiration of five years from the date of grant.
Restricted Stock Unit Awards. Restricted stock units are granted under restricted stock unit award agreements adopted by the plan administrator. Restricted stock units may be granted in consideration for any form of legal consideration that may be acceptable to our board of directors and permissible under applicable law. A restricted stock unit may be settled by cash, delivery of stock, a combination of cash and stock as deemed appropriate by the plan administrator, or in any other form of consideration set forth in the restricted stock unit agreement. Additionally, dividend equivalents may be credited in respect of shares covered by a restricted stock unit. Except as
168
Table of Contents
otherwise provided in the applicable award agreement, restricted stock units that have not vested will be forfeited once the participant’s continuous service ends for any reason.
Restricted Stock Awards. Restricted stock awards are granted under restricted stock award agreements adopted by the plan administrator. A restricted stock award may be awarded in consideration for cash, check, bank draft or money order, past services to us, or any other form of legal consideration that may be acceptable to our board of directors and permissible under applicable law. The plan administrator determines the terms and conditions of restricted stock awards, including vesting and forfeiture terms. If a participant’s service relationship with us ends for any reason, we may receive any or all of the shares of our common stock held by the participant that have not vested as of the date the participant terminates service with us through a forfeiture condition or a repurchase right.
Stock Appreciation Rights. Stock appreciation rights are granted under stock appreciation grant agreements adopted by the plan administrator. The plan administrator determines the purchase price or strike price for a stock appreciation right, which generally cannot be less than 100% of the fair market value of our common stock on the date of grant. A stock appreciation right granted under the 2026 Plan vests at the rate specified in the stock appreciation right agreement as determined by the plan administrator.
Performance Awards. The 2026 Plan permits the grant of performance-based stock and cash awards. The plan administrator may structure awards so that the shares of our stock, cash, or other property will be issued or paid only following the achievement of certain pre-established performance goals during a designated performance period. The performance criteria that will be used to establish such performance goals may be based on any one of, or combination of, the following as determined by the plan administrator: earnings (including earnings per share and net earnings); earnings before interest, taxes and depreciation; earnings before interest, taxes, depreciation and amortization; total stockholder return; return on equity or average stockholder’s equity; return on assets, investment, or capital employed; share price; margin (including gross margin); income (before or after taxes); operating income; operating income after taxes; pre-tax profit; operating cash flow; sales or revenue targets; increases in revenue or product revenue; expenses and cost reduction goals; improvement in or attainment of working capital levels; economic value added (or an equivalent metric); market share; cash flow; cash flow per share; share price performance; debt reduction; customer satisfaction; stockholder’s equity; capital expenditures; debt levels; operating profit or net operating profit; growth of net income or operating income; billings; financing; regulatory milestones; stockholder liquidity; corporate governance and compliance; intellectual property; personnel matters; progress of internal research; progress of partnered programs; partner satisfaction; budget management; partner or collaborator achievements; internal controls, including those related to the Sarbanes-Oxley Act of 2002; investor relations, analysts and communication; implementation or completion of projects or processes; employee retention; number of users, including unique users; strategic partnerships or transactions (including in-licensing and out-licensing of intellectual property); establishing relationships with respect to the marketing, distribution and sale of our products; supply chain achievements; co-development, co-marketing, profit sharing, joint venture or other similar arrangements; individual performance goals; corporate development and planning goals; and other measures of performance selected by the plan administrator.
The performance goals may be based on a company-wide basis, with respect to one or more business units, divisions, affiliates, or business segments, and in either absolute terms or relative to the performance of one or more comparable companies or the performance of one or more relevant indices. Unless specified otherwise (i) in the award agreement at the time the award is granted or (ii) in such other document setting forth the performance goals at the time the goals are established, we will appropriately make adjustments in the method of calculating the attainment of performance goals as follows: (1) to exclude restructuring and/or other nonrecurring charges; (2) to exclude exchange rate effects; (3) to exclude the effects of changes to generally accepted accounting principles; (4) to exclude the effects of any statutory adjustments to corporate tax rates; (5) to exclude the effects of items that are “unusual” in nature or occur “infrequently” as determined under generally accepted accounting principles; (6) to exclude the dilutive effects of acquisitions or joint ventures; (7) to assume that any business divested by us achieved performance objectives at targeted levels during the balance of a performance period following such divestiture; (8) to exclude the effect of any change in the outstanding shares of our common stock by reason of any stock dividend or split, stock repurchase, reorganization, recapitalization, merger, consolidation, spin-off, combination or exchange of shares or other similar corporate change, or any distributions to common stockholders other than regular
169
Table of Contents
cash dividends; (9) to exclude the effects of stock based compensation and the award of bonuses under our bonus plans; (10) to exclude costs incurred in connection with potential acquisitions or divestitures that are required to be expensed under generally accepted accounting principles; and (11) to exclude the goodwill and intangible asset impairment charges that are required to be recorded under generally accepted accounting principles. In addition, we retain the discretion to reduce or eliminate the compensation or economic benefit due upon attainment of the goals. The performance goals may differ from participant to participant and from award to award.
Other Stock Awards. The plan administrator may grant other awards based in whole or in part by reference to our common stock. The plan administrator will set the number of shares under the stock award and all other terms and conditions of such awards.
Non-employee Director Compensation Limit. The aggregate value of all compensation granted or paid to any non-employee director with respect to any fiscal year following the year in which the underwriting agreement for this offering is executed, including stock awards granted and cash fees paid by us to such non-employee director, will not exceed $750,000 in total value, or in the event such non-employee director is first appointed or elected to the board of directors during such fiscal year, $1,000,000 in total value (in each case, calculating the value of any such stock awards based on the grant date fair value of such stock awards for financial reporting purposes).
Changes to Capital Structure. In the event there is a specified type of change in our capital structure, such as a stock split, reverse stock split, or recapitalization, appropriate adjustments will be made to (i) the class and maximum number of shares reserved for issuance under the 2026 Plan; (ii) the class and maximum number of shares that may be issued on the exercise of ISOs; and (iii) the class and number of shares and exercise price, strike price, or purchase price, if applicable, of all outstanding stock awards.
Corporate Transactions. The following applies to stock awards under the 2026 Plan in the event of a corporate transaction, unless otherwise provided in a participant’s stock award agreement or other written agreement with us or one of our affiliates or unless otherwise expressly provided by the plan administrator at the time of grant.
In the event of a corporate transaction (as defined in the 2026 Plan), any stock awards outstanding under the 2026 Plan may be assumed, continued or substituted for by any surviving or acquiring corporation (or its parent company), and any reacquisition or repurchase rights held by us with respect to the stock award may be assigned to the successor (or its parent company). If the surviving or acquiring corporation (or its parent company) does not assume, continue or substitute for such stock awards, then with respect to any such stock awards that are held by participants whose continuous service has not terminated prior to the effective time of the transaction, or current participants, the vesting (and exercisability, if applicable) of such stock awards will be accelerated in full to a date prior to the effective time of the transaction (contingent upon the effectiveness of the transaction), and such stock awards will terminate if not exercised (if applicable) at or prior to the effective time of the transaction, and any reacquisition or repurchase rights held by us with respect to such stock awards will lapse (contingent upon the effectiveness of the transaction). With respect to performance awards with multiple vesting levels depending on performance level, unless otherwise provided by an award agreement or by the plan administrator, the award will accelerate at 100% of target. If the surviving or acquiring corporation (or its parent company) does not assume, continue or substitute for such stock awards, then with respect to any such stock awards that are held by persons other than current participants, such awards will terminate if not exercised (if applicable) prior to the effective time of the transaction, except that any reacquisition or repurchase rights held by us with respect to such stock awards will not terminate and may continue to be exercised notwithstanding the transaction. The plan administrator is not obligated to treat all stock awards or portions of stock awards in the same manner and is not obligated to take the same actions with respect to all participants.
In the event a stock award will terminate if not exercised prior to the effective time of a transaction, the plan administrator may provide, in its sole discretion, that the holder of such stock award may not exercise such stock award but instead will receive a payment equal in value to the excess (if any) of (i) the value of the property the participant would have received upon the exercise of the stock award over (ii) any exercise price payable by such holder in connection with such exercise.
Under our 2026 Plan, a corporate transaction is defined to include: (i) a sale of all or substantially all of our assets; (ii) the sale or disposition of at least 50% of our outstanding securities; (iii) the consummation of a merger or
170
Table of Contents
consolidation where we do not survive the transaction; and (iv) the consummation of a merger or consolidation where we do survive the transaction but the shares of our common stock outstanding before such transaction are converted or exchanged into other property by virtue of the transaction, unless otherwise provided in an award agreement or other written agreement between us and the award holder.
Change in Control. In the event of a change in control, as defined under our 2026 Plan, awards granted under our 2026 Plan will not receive automatic acceleration of vesting and exercisability, although this treatment may be provided for in an award agreement.
Under the 2026 Plan, a change in control is defined to include (i) the acquisition by any person or company of more than 50% of the combined voting power of our then outstanding stock; (ii) a merger, consolidation or similar transaction in which our stockholders immediately before the transaction do not own, directly or indirectly, more than 50% of the combined voting power of the surviving entity (or the parent of the surviving entity); (iii) a sale, lease, exclusive license or other disposition of all or substantially all of our assets other than to an entity more than 50% of the combined voting power of which is owned by our stockholders; and (iv) an unapproved change in the majority of the board of directors.
Clawback. All awards granted under the 2026 Plan will be subject to recoupment in accordance with any clawback policy that we are required to adopt pursuant to the listing standards of any national securities exchange or association on which our securities are listed or as is otherwise required by the U.S. Dodd-Frank Wall Street Reform and Consumer Protection Act or other applicable law. In addition, our board of directors may impose such other clawback, recovery or recoupment provisions in a stock award agreement as our board of directors determines necessary or appropriate.
Transferability. A participant may not transfer stock awards under our 2026 Plan other than by will, the laws of descent and distribution, or as otherwise provided under our 2026 Plan.
Plan Amendment or Termination. Our board of directors has the authority to amend, suspend, or terminate our 2026 Plan, provided that such action does not materially impair the existing rights of any participant without such participant’s written consent. Certain material amendments also require the approval of our stockholders. No ISOs may be granted after the tenth anniversary of the date our board of directors adopted our 2026 Plan. No stock awards may be granted under our 2026 Plan while it is suspended or after it is terminated.
2026 Employee Stock Purchase Plan
Our board of directors adopted our ESPP in August 2026 and our stockholders approved our ESPP in September 2026. The ESPP will become effective upon the execution of the underwriting agreement for this offering. The purpose of the ESPP is to secure and retain the services of new employees, to retain the services of existing employees, and to provide incentives for such individuals to exert maximum efforts toward our success and that of our affiliates. Our ESPP will include two components. One component will be designed to allow eligible U.S. employees to purchase our shares of our common stock in a manner that may qualify for favorable tax treatment under Section 423 of the Code. The other component will permit the grant of purchase rights that do not qualify for such favorable tax treatment in order to allow deviations necessary to permit participation by eligible employees who are foreign nationals or employed outside of the United States while complying with applicable foreign laws.
Share Reserve. Following this offering, the ESPP authorizes the issuance of 611,682 shares of our common stock under purchase rights granted to our employees or to employees of any of our designated affiliates. The number of shares of our common stock reserved for issuance will automatically increase on January 1 of each calendar year, beginning on January 1, 2027 (assuming the ESPP becomes effective in 2026) through January 1, 2036, by the lesser of (a) 1.0% of the total number of shares of our common stock (including shares issuable upon exercise, conversion, or exchange of all then-outstanding prefunded warrants, or other rights exercisable for or convertible into, directly or indirectly, shares of common stock (whether vested or unvested), but excluding shares issuable upon the exercise of then-outstanding options and the shares reserved and available for issuance under our then-existing equity incentive plan(s) and employee stock purchase plan(s), if any) outstanding on December 31 of the immediately preceding calendar year; and (b) 1,835,046 shares; provided that before the date of any such increase, our board of directors may determine that such increase will be less than the amount set forth in clauses (a) and (b). As of the date hereof, no shares of our common stock have been purchased under the ESPP.
171
Table of Contents
Administration. Our board of directors, or a duly authorized committee thereof, will administer our ESPP. Our board of directors may delegate concurrent authority to administer the ESPP to our compensation committee under the terms of the compensation committee’s charter. The ESPP is implemented through a series of offerings under which eligible employees are granted purchase rights to purchase shares of our common stock on specified dates during such offerings. Under the ESPP, we may specify offerings with durations of not more than 27 months and may specify shorter purchase periods within each offering. Each offering will have one or more purchase dates on which shares of our common stock will be purchased for employees participating in the offering. An offering under the ESPP may be terminated under certain circumstances.
Payroll Deductions. Generally, all regular employees, including executive officers, employed by us or by any of our designated affiliates, may participate in the ESPP and may contribute, normally through payroll deductions, up to 15% of their earnings (as defined in the ESPP) for the purchase of our common stock under the ESPP. Unless otherwise determined by our board of directors, common stock will be purchased for the accounts of employees participating in the ESPP at a price per share that is at least the lesser of (i) 85% of the fair market value of a share of our common stock on the first date of an offering; or (ii) 85% of the fair market value of a share of our common stock on the date of purchase.
Limitations. Employees may have to satisfy one or more of the following service requirements before participating in the ESPP, as determined by our board of directors, including: (i) customary employment with us or one of our affiliates for more than 20 hours per week and more than five months per calendar year; or (ii) continuous employment with us or one of our affiliates for a minimum period of time (not to exceed two years). No employee may purchase shares under the ESPP at a rate in excess of $25,000 worth of our common stock based on the fair market value per share of our common stock at the beginning of an offering for each year such a purchase right is outstanding. Finally, no employee will be eligible for the grant of any purchase rights under the ESPP if immediately after such rights are granted, such employee has voting power over 5% or more of our outstanding capital stock measured by vote or value under Section 424(d) of the Code.
Changes to Capital Structure. In the event that there occurs a change in our capital structure through such actions as a stock split, merger, consolidation, reorganization, recapitalization, reincorporation, stock dividend, dividend in property other than cash, large nonrecurring cash dividend, liquidating dividend, combination of shares, exchange of shares, change in corporate structure, or similar transaction, the board of directors will make appropriate adjustments to: (i) the number of shares reserved under the ESPP; (ii) the maximum number of shares by which the share reserve may increase automatically each year; (iii) the number of shares and purchase price of all outstanding purchase rights; and (iv) the number of shares that are subject to purchase limits under ongoing offerings.
Corporate Transactions. In the event of certain significant corporate transactions (as defined in the ESPP), including: (i) a sale of all or substantially all of our assets; (ii) the sale or disposition of more than 50% of our outstanding securities; (iii) the consummation of a merger or consolidation where we do not survive the transaction; and (iv) the consummation of a merger or consolidation where we do survive the transaction but the shares of our common stock outstanding immediately before such transaction are converted or exchanged into other property by virtue of the transaction, any then-outstanding rights to purchase our stock under the ESPP may be assumed, continued or substituted for by any surviving or acquiring entity (or its parent company). If the surviving or acquiring entity (or its parent company) elects not to assume, continue, or substitute for such purchase rights, then the participants’ accumulated payroll contributions will be used to purchase shares of our common stock within ten business days before such corporate transaction, and such purchase rights will terminate immediately.
ESPP Amendment or Termination. Our board of directors has the authority to amend or terminate our ESPP, provided that except in certain circumstances such amendment or termination may not materially impair any outstanding purchase rights without the holder’s consent. We will obtain stockholder approval of any amendment to our ESPP as required by applicable law or listing requirements.
2022 Stock Plan
Our board of directors adopted, and our stockholders approved, our 2022 Plan in February 2022. Our 2022 Plan was most recently amended in August 2026, which amendment will be effective immediately prior to the effectiveness of this registration statement. The 2022 Plan will terminate on the date the 2026 Plan becomes effective, and thereafter no further stock awards will be granted under the 2022 Plan. However, any outstanding
172
Table of Contents
stock awards granted under the 2022 Plan will remain outstanding, subject to the terms of our 2022 Plan and award agreements, until such outstanding stock awards are exercised or until such outstanding stock awards terminate or expire by their terms.
Types of Awards. Our 2022 Plan provides for the grant of ISOs to employees, including employees of our affiliates, and for the grant of NSOs, stock awards and restricted stock units to employees, directors, and consultants, including employees and consultants of our affiliates.
Share Reserve. Subject to certain capitalization adjustments, the common stock that may be issued pursuant to stock awards under the 2022 Plan will not exceed in the aggregate 6,196,109 shares of our common stock. The shares of our common stock subject to the 2022 Plan may be unissued shares or reacquired shares, bought on the market or otherwise. As of December 31, 2025 and June 30, 2026, 3,367,661 and 464,837 shares of our common stock, respectively, remained available for future issuance under the 2022 Plan.
Administration. Our board of directors or one or more duly authorized committees of our board of directors (referred to herein as the plan administrator) administers our 2022 Plan and the stock awards granted under it. Under our 2022 Plan, the plan administrator has full authority and discretion to take any actions it deems necessary or advisable for the administration of the 2022 Plan.
Options. Each option granted under the 2022 Plan is in such form and contains such terms and conditions as determined by the plan administrator. The exercise price of an option under the 2022 Plan generally cannot be less than 100% of the fair market value of our common stock on the date of grant (or 110% of the fair market value for ISOs granted to certain major stockholders).
The purchase price of shares of our common stock acquired pursuant to an option under the 2022 Plan may be paid in cash or cash equivalents. In addition, the plan administrator may permit payment through any of the following methods: (i) via promissory note, (ii) by surrender of shares of our common stock, (iii) by a cashless exercise arrangement established by us, (iv) by a net exercise arrangement, or (v) any other form permitted by the Delaware General Corporation Law and as provided in the applicable award agreement.
ISOs are not transferable except by will or by the laws of descent and distribution and are exercisable during the lifetime of the optionholder only by the optionholder. NSOs generally are transferable to the extent provided in the option agreement to the extent permitted by Rule 701 under the Securities Act of 1933, as amended (Securities Act).
In the event that an optionholder’s continuous service terminates (other than upon the optionholder’s death or disability), the optionholder may exercise his or her option (to the extent that the optionholder was entitled to exercise such option as of the date of termination) but only within such period of time ending on the earlier of (i) the date three months following the termination of the optionholder’s continuous service (or such earlier or later period as the plan administrator may determine but in no event earlier than thirty days after optionholder’s termination of service), or (ii) the expiration of the term of the option as set forth in the option agreement. In the event of the optionholder’s termination as a result of disability, the optionholder may exercise his or her option on the earliest of (i) the date six months following the termination of optionholder’s continuous service (or such later date as the plan administrator may determine), or (ii) the expiration term of the option as set forth in the option agreement. If an optionholder dies while they are in service, the optionholder’s options will expire on the earlier of (i) the date twelve months following the optionholder’s death (or such earlier or later date as the plan administrator may determine but in no event earlier than six months after the optionholder’s death), or (ii) the expiration term of the option as set forth in the option agreement. If, after termination, the optionholder does not exercise his or her option within the time specified in the option agreement, the option will terminate.
The plan administrator may grant options that can be exercised before the shares subject to the option have vested. If a participant exercises unvested shares subject to an option, the participant will receive unvested (i.e. restricted) shares subject to a right of repurchase in favor of the company that will lapse over the original vesting schedule for the option while the participant remains in continuous service. If a participant’s service relationship with us ends for any reason, we may receive any or all of the shares of our common stock held by the participant that have not vested as of the date the participant terminates service with us through a forfeiture condition or a repurchase right.
Corporate Transactions. Effective upon the amendment of the 2022 Plan as of immediately prior to the effectiveness of this registration statement, the following will apply to stock awards under the 2022 Plan in the event of a
173
Table of Contents
corporate transaction, unless otherwise provided in a participant’s stock award agreement or other written agreement with us or one of our affiliates or unless otherwise expressly provided by the plan administrator at the time of grant.
In the event of a corporate transaction (as defined in the 2022 Plan), any stock awards outstanding under the 2022 Plan may be assumed, continued or substituted for by any surviving or acquiring corporation (or its parent company), and any reacquisition or repurchase rights held by us with respect to the stock award may be assigned to the successor (or its parent company). If the surviving or acquiring corporation (or its parent company) does not assume, continue or substitute for such stock awards, then with respect to any such stock awards that are held by participants whose continuous service has not terminated prior to the effective time of the transaction, or current participants, the vesting (and exercisability, if applicable) of such stock awards will be accelerated in full to a date prior to the effective time of the transaction (contingent upon the effectiveness of the transaction), and such stock awards will terminate if not exercised (if applicable) at or prior to the effective time of the transaction, and any reacquisition or repurchase rights held by us with respect to such stock awards will lapse (contingent upon the effectiveness of the transaction). If the surviving or acquiring corporation (or its parent company) does not assume, continue or substitute for such stock awards, then with respect to any such stock awards that are held by persons other than current participants, such awards will terminate if not exercised (if applicable) prior to the effective time of the transaction, except that any reacquisition or repurchase rights held by us with respect to such stock awards will not terminate and may continue to be exercised notwithstanding the transaction. The plan administrator is not obligated to treat all stock awards or portions of stock awards in the same manner and is not obligated to take the same actions with respect to all participants.
In the event a stock award will terminate if not exercised prior to the effective time of a transaction, the plan administrator may provide, in its sole discretion, that the holder of such stock award may not exercise such stock award but instead will receive a payment equal in value to the excess (if any) of (i) the value of the property the participant would have received upon the exercise of the stock award over (ii) any exercise price payable by such holder in connection with such exercise.
In addition, with respect to options, the plan administrator may (A) suspend the participant’s right to exercise the option during a limited period of time preceding the closing of the Corporate Transaction if such suspension is administratively necessary to facilitate the closing of the Corporate Transaction and/ or (B) terminate any right the participant has to early exercise the option, such that following the closing of the Corporate Transaction, the option may only be exercised to the extent it is vested.
Capitalization Adjustments. In the event of certain changes in our capitalization, the number and class of securities subject to the 2022 Plan, and the stock awards outstanding under the 2022 Plan and any applicable exercise price will be appropriately adjusted.
Plan Amendment or Termination. Our board of directors may amend, suspend, or terminate the 2022 Plan at any time. The 2022 Plan will terminate on the date the 2026 Plan becomes effective, and thereafter no further stock awards will be granted under the 2022 Plan.
Electra Therapeutics LLC 2023 Equity Incentive Plan
The Electra LLC board of managers adopted the 2023 Plan in June 2023, pursuant to which common units of Electra LLC intended to qualify as “profits interests” for U.S. federal income tax purposes (Profits Interest Units) have been granted. As of December 31, 2025 and June 30, 2026, there were 9,813,409 Profits Interest Units outstanding under the 2023 Plan. As discussed further above, prior to the closing of this offering, Electra LLC will effect the LLC Distribution, pursuant to which Electra LLC will distribute shares of our Series A redeemable convertible preferred stock to holders of outstanding Profits Interest Units. Any shares of Series A redeemable convertible preferred stock received in the LLC Distribution will automatically convert into shares of our common stock immediately prior to the closing of this offering.
Authorized Units. Subject to certain capitalization adjustments, the aggregate number of common units that may be issued pursuant to all awards under the 2023 Plan is 9,930,785 common units. Any common units subject to awards that are forfeited or canceled are thereafter available for issuance under the 2023 Plan.
Type of Awards. The 2023 Plan provides for the grant of Profits Interest Units, which are interests in Electra LLC intended to qualify as “profits interests” for U.S. federal income tax purposes, and awards of common units to
174
Table of Contents
certain employees, advisors, consultants, and service providers of Electra LLC. To achieve the tax treatment with respect to the Profits Interest Units, each Profits Interest Unit is assigned a “distribution threshold” to result in a liquidation value of $0.
Plan Administration. Electra LLC’s board of managers administers and interprets the 2023 Plan. Under the 2023 Plan, the board of managers has the full authority and discretion to take any actions it deems necessary or advisable for the administration of the 2023 Plan.
Adjustments to Common Units. In the event of a subdivision, reclassification, or combination of units, or other change in Electra LLC’s capital structure, the administrator shall make appropriate adjustments to the number of common units available for grant under the 2023 Plan and to the distribution threshold and number of common units subject to awards then outstanding.
Corporate Transactions. In the event Electra LLC consummates a Liquidation Event or Incorporation (each, as defined in the LLC Agreement), all Profits Interest Units will be treated as set forth in the LLC Agreement and the definitive transaction agreement, which need not treat all awards in an identical manner, and may include one or more of the following: (1) continuation of the Profits Interest Units if Electra LLC is the surviving entity, (2) cancellation of the Profits Interest Units in exchange for payment equal to the fair market value of the common units as of the transaction to the extent vested, (3) conversion of the Profits Interest Units by the surviving entity or its parent into equity of the surviving entity or its parent, (4) an amount in cash or other consideration from a sale of assets of Electra LLC constituting a Liquidation Event otherwise distributable to a common unit, reduced by the distribution threshold, (5) cancellation for no consideration if the distribution threshold exceeds the amount that would be distributed or payable in connection with the transaction, or if the common units would otherwise receive no distributions or proceeds in the transaction, and (6) the repurchase or forfeiture of any or all unvested Profits Interest Units.
Distributions. As further set forth in the LLC Agreement, holders of Profits Interest Units are entitled to participate in certain distributions above the distribution threshold set forth in their Profits Interest Unit agreements.
Transferability. Profits Interest Units granted under the 2023 Plan are subject to the restrictions on transfer in the LLC Agreement and any other transfer restrictions as determined by Electra LLC.
Plan Amendment or Termination. The administrator may amend or terminate the 2023 Plan at any time. Certain amendments or modifications may require member consent as provided in the LLC Agreement.
Director Compensation
Historically, we have not had a formal compensation policy with respect to service on our board of directors. However, we have entered into individual agreements with each of Mr. Fust and Drs. Rosenthal and Stagliano with respect to their services on our board and pursuant to which they are provided certain compensation, as described further below. We did not provide compensation to our other non-employee directors related to their board service during fiscal year ended December 31, 2025.
Dr. Dong is a member of our board of directors and an employee of the Company, but does not receive any additional compensation for service as a director. The compensation earned by or paid to Dr. Dong as a named executive officer for the fiscal year ended December 31, 2025 is set forth above in the subsection titled “—Summary Compensation Table for the Year Ended December 31, 2025.”
175
Table of Contents
| NAME |
FEES EARNED OR PAID IN CASH ($) (1) |
OPTION AWARDS ($) (2)(3) |
ALL OTHER COMPENSATION ($) |
TOTAL ($) |
||||||||||||
| Matthew Fust, M.B.A. |
45,000 | — | — | 45,000 | ||||||||||||
| Thomas Geninatti, Ph.D. (5) |
— | — | — | — | ||||||||||||
| Carl L. Gordon, Ph.D., C.F.A. |
— | — | — | — | ||||||||||||
| Adam Rosenthal, Ph.D. (4) |
19,795 | — | — | 19,795 | ||||||||||||
| Beth Seidenberg, M.D. (5) |
— | — | — | — | ||||||||||||
| Nancy Stagliano, Ph.D. |
100,000 | — | — | 100,000 | ||||||||||||
| (1) | Represents fees paid in fiscal year 2025. Dr. Geninatti, Dr. Gordon and Dr. Seidenberg did not receive compensation for their service to us as non-employee directors in fiscal year 2025. |
| (2) | As of December 31, 2025, the aggregate number of shares underlying outstanding options to purchase our common stock held by our non-employee directors was 59,086, 103,401 and 94,538 for each of Mr. Fust, Dr. Rosenthal and Dr. Stagliano, respectively. None of our other non-employee directors held any outstanding options as of December 31, 2025. |
| (3) | As of December 31, 2025, the aggregate number of Profits Interest Units of Electra LLC outstanding held by our non-employee directors was 671,828 and 1,080,348 for each of Dr. Rosenthal and Dr. Stagliano, respectively. In addition, as of December 31, 2025, Dr. Rosenthal held an aggregate of 5,000,000 fully vested Common Units of Electra LLC. |
| (4) | Dr. Rosenthal resigned from our board of directors in October 2025, and the cash compensation included in the table above reflects his cash compensation earned through October 2025. |
| (5) | Drs. Geninatti and Seidenberg have notified us that they intend to resign from our board of directors effective as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. |
We reimburse all of our non-employee directors for their reasonable out-of-pocket expenses incurred in attending board of directors and committee meetings.
In October 2023, we entered into an individual agreement with Mr. Fust, which was subsequently amended in March 2024, with respect to his services on our board of directors. Pursuant to Mr. Fust’s agreement, he is eligible to receive an annual cash retainer of $25,000 for his services on our board of directors, plus an additional $10,000 for his services on the audit committee of the board of directors and an additional $10,000 for his services on the compensation committee of our board of directors. Mr. Fust’s agreement further provides that we will reimburse Mr. Fust for reasonable expenses incurred in connection with attendance at meetings of the board or committees of the board.
In November 2023, we entered into an individual agreement with Dr. Rosenthal, which was subsequently amended in October 2025, with respect to his services on our board of directors. Pursuant to Dr. Rosenthal’s agreement, he was eligible to receive an annual cash retainer of $25,000 for his services on our board of directors. In addition, Dr. Rosenthal’s agreement provides that we will reimburse Dr. Rosenthal for reasonable expenses incurred in connection with attendance at meetings of the board or committees of the board. Dr. Rosenthal resigned from our board of directors in October 2025 and his cash compensation for 2025 was prorated for the time he served on our board of directors during the year. In connection with his resignation, Dr. Rosenthal is no longer entitled to any compensation under this agreement.
In January 2024, we entered into an individual agreement with Dr. Stagliano with respect to her services as the Chairperson of our board of directors. Pursuant to Dr. Stagliano’s agreement, she is eligible to receive an annual cash retainer of $100,000 for her services as the Chairperson of our board of directors. In addition, Dr. Stagliano’s agreement provides that we will reimburse Dr. Stagliano for reasonable expenses incurred in connection with attendance at meetings of the board or committees of the board.
IPO Grants
At the time of the effectiveness of the 2026 Plan, our board of directors will grant options to purchase shares of our common stock to our non-employee directors, with an exercise price equal to the initial public offering price per share. As of such date, each of Mr. Fust and Drs. Gordon and Stagliano will be granted options to purchase 22,632, 22,632, and 68,819 shares of our common stock, respectively, with the number of shares subject to Dr. Stagliano’s option reflecting an additional 46,187 shares in recognition of her prior and continued board of directors’ service to
176
Table of Contents
the Company, including as the Chairperson of our board of directors and the efforts she contributed toward completion of this offering in her role as a member of our board of directors. The shares underlying the options vest on the earlier of (i) the date that is 12 months following the grant date or (ii) the date of our next annual stockholder meeting.
Post-IPO Director Compensation Policy
Our board of directors adopted a non-employee director compensation policy in August 2026 (the Post-IPO Compensation Policy) that will become effective upon the execution and delivery of the underwriting agreement related to this offering and will be applicable to all of our non-employee directors. We will terminate the individual agreements with each of Dr. Stagliano and Mr. Fust described above effective and contingent upon the effectiveness of the non-employee director compensation policy. The Post-IPO Compensation Policy provides that each such non-employee director will receive the following compensation for service on our board of directors:
| ∎ | an annual cash retainer of $40,000; |
| ∎ | an additional annual cash retainer of $30,000 for service as the non-executive chair of the board of directors; |
| ∎ | an additional annual cash retainer of $10,000, $7,500 and $5,000 for service as a non-chair member of the audit committee, compensation committee, and the nominating and corporate governance committee, respectively; and |
| ∎ | an additional annual cash retainer of $20,000, $15,000 and $10,000 for service as chair of the audit committee, chair of the compensation committee, and chair of the nominating and corporate governance committee, respectively (in lieu of the non-chair committee member retainer above). |
Each of the annual cash retainers will be paid in equal quarterly installments, payable in arrears on the last day of each fiscal quarter in which the service occurred. If a non-employee director joins our board of directors or a committee of our board of directors at a time other than effective as of the first day of a fiscal quarter, each annual cash retainer described above will be pro-rated on days served in the applicable fiscal quarter, with the pro-rated amount paid for the first fiscal quarter in which such non-employee director provides the service and regular full quarterly payments thereafter. All annual cash retainers are vested upon payment.
In addition, the Post-IPO Compensation Policy provides that each such non-employee director will be eligible to receive the following option grants for service on our board of directors and committees of our board of directors:
| ∎ | an initial option grant to purchase 22,632 shares of our common stock, vesting in a series of 36 equal monthly installments; and |
| ∎ | an annual option grant to purchase 45,264 shares of our common stock, vesting on the earlier of (i) the date that is 12 months following the grant date or (ii) the date of our next annual stockholder meeting. |
Each of the option grants described above will be granted under our 2026 Plan, the terms of which are described in more detail above under “—Equity Benefit Plans—2026 Equity Incentive Plan.” Each such option grant will vest and become exercisable subject to the director’s continuous service with us, provided that each option will vest in full upon a “change in control” (as defined in the 2026 Plan) of the company, subject to the director’s continuous service through immediately prior to the change in control. The term of each option will be ten years, subject to earlier termination as provided in the 2026 Plan.
We will reimburse all of our non-employee directors for their reasonable out-of-pocket expenses incurred in attending board of directors and committee meetings and other reasonable expenses incurred in connection with the non-employee director’s service on the board, provided that each such non-employee director submit appropriate documentation substantiating such expenses to us and in accordance with our travel and expense policy.
Limitations of Liability and Indemnification Matters
Our amended and restated certificate of incorporation that will become effective immediately prior to the closing of this offering limits the liability of our current and former directors and officers for monetary damages to the fullest
177
Table of Contents
extent permitted by Delaware law. Delaware law provides that directors and officers of a corporation will not be personally liable for monetary damages for any breach of fiduciary duties as directors or officers, except liability for:
| ∎ | any breach of the director’s or officer’s duty of loyalty to the corporation or its stockholders; |
| ∎ | any act or omission not in good faith or that involves intentional misconduct or a knowing violation of law; |
| ∎ | as a director, unlawful payments of dividends or unlawful stock repurchases or redemptions; |
| ∎ | as an officer, derivative claims brought on behalf of the corporation by a stockholder; or |
| ∎ | any transaction from which the director or officer derived an improper personal benefit. |
This limitation of liability does not apply to liabilities arising under federal securities laws and does not affect the availability of equitable remedies such as injunctive relief or rescission.
Our amended and restated certificate of incorporation will authorize us to indemnify our directors, officers, employees, and other agents to the fullest extent permitted by Delaware law. Our amended and restated bylaws will provide that we are required to indemnify our directors and officers to the fullest extent permitted by Delaware law and may indemnify our other employees and agents. Our amended and restated bylaws will also provide that, on satisfaction of certain conditions, we will advance expenses incurred by a director or officer in advance of the final disposition of any action or proceeding, and permit us to secure insurance on behalf of any officer, director, employee, or other agent for any liability arising out of his or her actions in that capacity regardless of whether we would otherwise be permitted to indemnify him or her under the provisions of Delaware law. We have entered and expect to continue to enter into agreements to indemnify our directors, executive officers, and other employees as determined by the board of directors. With certain exceptions, these agreements provide for indemnification for related expenses including attorneys’ fees, judgments, fines, and settlement amounts incurred by any of these individuals in any action or proceeding.
We believe that the amended and restated certificate of incorporation, amended and restated bylaw provisions and indemnification agreements are necessary to attract and retain qualified persons as directors and officers. We also maintain customary directors’ and officers’ liability insurance.
The limitation of liability and indemnification provisions in our amended and restated certificate of incorporation and amended and restated bylaws may discourage stockholders from bringing a lawsuit against our directors for breach of their fiduciary duty. They may also reduce the likelihood of derivative litigation against our directors and officers, even though an action, if successful, might benefit us and other stockholders. Further, a stockholder’s investment may be adversely affected to the extent that we pay the costs of settlement and damage awards against directors and officers as required by these indemnification provisions.
Insofar as indemnification for liabilities arising under the Securities Act may be permitted for directors, executive officers, or persons controlling us, we have been informed that, in the opinion of the SEC, such indemnification is against public policy as expressed in the Securities Act and is therefore unenforceable.
At present, there is no pending litigation or proceeding involving any of our directors, executive officers or employees for which indemnification is sought, and we are not aware of any threatened litigation that may result in claims for indemnification.
Rule 10b5-1 Sales Plans
Our directors and executive officers may adopt written plans, known as Rule 10b5-1 plans, in which they will contract with a broker to buy or sell shares of our common stock on a periodic basis. Under a Rule 10b5-1 plan, a broker executes trades pursuant to parameters established by the director or executive officer when entering into the plan, without further direction from them. The director or executive officer may amend a Rule 10b5-1 plan in some circumstances and may terminate a Rule 10b5-1 plan at any time. Our directors and executive officers also may buy or sell additional shares outside of a Rule 10b5-1 plan when they are not in possession of material nonpublic information subject to compliance with the terms of our insider trading policy and any applicable Rule 10b5-1 guidelines.
178
Table of Contents
CERTAIN RELATIONSHIPS AND RELATED PERSON TRANSACTIONS
The following includes a summary of transactions since January 1, 2023, to which we have been a party, in which the amount involved in the transaction exceeded the lesser of $120,000 or 1% of the average of our total assets as of December 31, 2024 and 2025, and in which any of our directors, executive officers, or, to our knowledge, beneficial owners of more than 5% of our capital stock at the time of such transaction, or any member of the immediate family of any of the foregoing persons had or will have a direct or indirect material interest, other than equity and other compensation, termination, change in control, and other arrangements, which are described in the section titled “Executive and Director Compensation.”
Convertible Promissory Note Financing
In March 2025 and June 2025, we issued and sold to investors unsecured convertible promissory notes in an aggregate principal amount of $40.0 million of convertible promissory notes (the Convertible Notes). The Convertible Notes bore interest at a rate of 8.0% per annum. In connection with the initial closing of our Series C Financing (as defined below) in October 2025, the aggregate principal amount and accrued interest on the convertible promissory notes were converted into 4,080,265 shares of our Series C redeemable convertible preferred stock at an aggregate conversion price of $41.4 million.
The following table sets forth the aggregate principal amount of the Convertible Notes purchased by holders of more than 5% of our capital stock and entities affiliated with our directors and executive officers.
|
PARTICIPANTS (1) |
CONVERTIBLE PROMISSORY NOTES PURCHASED ($) |
|||
| OrbiMed Private Investments VII, LP |
7,044,002 | |||
| Redmile Biopharma Investments II, L.P. |
7,050,871 | |||
| Westlake BioPartners Fund II, L.P. |
11,720,022 | |||
| (1) | Additional details regarding these stockholders and their equity holdings are provided in this prospectus under the section titled “Principal Stockholders” and below in the subsection titled “—Series C Redeemable Convertible Preferred Stock Financing.” |
Series C Redeemable Convertible Preferred Stock Financing
In October 2025 and June 2026, we issued and sold to investors, in our Series C redeemable convertible preferred stock financing (the Series C Financing), an aggregate of 18,133,057 shares of our Series C redeemable convertible preferred stock at a purchase price of $10.15 per share for aggregate cash proceeds of approximately $182.7 million, including the conversion of the principal and accrued interest under the Convertible Notes.
The following table summarizes the shares of Series C redeemable convertible preferred stock issued in our Series C Financing to holders of more than 5% of our capital stock and entities affiliated with our directors and executive officers.
| PARTICIPANTS |
SERIES C CONVERTIBLE PREFERRED STOCK |
AGGREGATE PURCHASE PRICE ($) |
||||||
| Aventis Inc. (1) |
2,461,948 | 24,999,994 | ||||||
| LSP 7 Coöperatief U.A. (2) |
2,461,948 | 24,999,994 | ||||||
| Entities affiliated with OrbiMed Private Investments VII, LP (3) |
2,461,593 | 24,996,414 | ||||||
| Redmile Biopharma Investments II, L.P. (4) |
1,704,015 | 17,303,530 | ||||||
| Westlake BioPartners Fund II, L.P. (5) |
1,195,523 | 12,140,012 | ||||||
| Nextech VIII SCSp (6) |
1,969,559 | 19,999,997 | ||||||
179
Table of Contents
| (1) | Aventis Inc. (Sanofi) holds more than 5% of our outstanding capital stock. |
| (2) | LSP 7 Coöperatief U.A. (EQT) holds more than 5% of our outstanding capital stock. |
| (3) | Consists of (i) 1,703,314 shares of Series C redeemable convertible preferred stock held by OPI VII and (ii) 758,279 shares of Series C redeemable convertible preferred stock held by OrbiMed Genesis Master Fund, LP (Orbi Genesis and together with OPI VII, OrbiMed). OrbiMed holds more than 5% of our outstanding capital stock. OrbiMed Capital GP VII LLC (GP VII) is the general partner of OPI VII. OrbiMed Genesis GP LLC (Genesis GP) is the general partner of Orbi Genesis and OrbiMed Advisors LLC (OrbiMed Advisors) is the managing member of GP VII and Genesis GP. By virtue of such relationships, GP VII, Genesis GP, and OrbiMed Advisors may be deemed to have voting power and investment power over the securities held by OPI VII and Orbi Genesis and as a result, may be deemed to have beneficial ownership of such securities. OrbiMed Advisors exercises voting and investment power through a management committee comprised of Dr. Gordon, Sven H. Borho, and W. Carter Neild, each of whom disclaims beneficial ownership of the shares held by OPI VII and Orbi Genesis. Dr. Gordon, a member of our board of directors, is a Founding Member and Managing Partner at OrbiMed Advisors. |
| (4) | Redmile Biopharma Investments II, L.P. (Redmile) holds more than 5% of our outstanding capital stock. |
| (5) | Westlake holds more than 5% of our outstanding capital stock. Dr. Seidenberg, a member of our board of directors, is the sole managing director of Westlake BioPartners GP II, LLC, the general partner of Westlake. Dr. Seidenberg has notified us that she intends to resign from our board of directors as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. |
| (6) | Dr. Geninatti, a member of our board of directors, is a Principal of Nextech Invest AG, which is affiliated with Nextech VIII SCSp. Dr. Geninatti has notified us that he intends to resign from our board of directors as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. |
Investor Agreements
In connection with our redeemable convertible preferred stock financings, we entered into investors’ rights, right of first refusal and co-sale, and voting agreements and management rights letters, which contain, among other things, registration rights, information rights, voting rights, rights of first refusal, and certain other rights and covenants, with certain holders of our capital stock, including entities affiliated with Sanofi, EQT, Nextech, OrbiMed, Redmile, Westlake, and certain of our other stockholders. These agreements will terminate upon the closing of this offering, except for the registration rights granted under our investor rights agreement (defined below), as more fully described in the section titled “Description of Capital Stock—Registration Rights,” certain rights granted to Sanofi under the Sanofi Side Letter, as defined and more fully described below in the subsection titled “—Sanofi Side Letter,” and other certain rights granted to Redmile under the Redmile MRL, as defined and more fully described below in the subsection titled “—Redmile Management Rights Letter.” See also the section titled “Principal Stockholders” for additional information regarding beneficial ownership of our capital stock.
Sanofi Side Letter
We entered into a side letter with Sanofi (the Sanofi Side Letter), a holder of more than 5% of our outstanding capital stock, in October 2025 in connection with Sanofi’s purchase of shares of Series C redeemable convertible preferred stock. Under the Sanofi Side Letter, we agreed, among other things: (i) to allow Sanofi to have a representative attend, as an observer, meetings of our scientific and clinical advisory boards and committees pertaining to some of our product candidates; (ii) to allow Sanofi to request update meetings and access to certain clinical and regulatory data relating to our product candidates; (iii) to provide Sanofi with written notice relating to certain potential transactions involving our product candidates and the ability to participate in any such process; and (iv) to have Sanofi’s restrictive legends on the shares of our capital stock removed. The rights described above will remain in effect until the earlier of (x) a Liquidation Event (as defined in our Amended and Restated Certificate of Incorporation as currently in effect), (y) if Sanofi becomes a sanctioned party, and (z) such time as Sanofi no longer satisfies certain ownership requirements with respect to the shares of our capital stock.
Redmile Management Rights Letter
We entered into an amended and restated management rights letter (the Redmile MRL) with Redmile, a holder of more than 5% of our outstanding capital stock, in October 2025 in connection with Redmile’s purchase of shares of Series C redeemable convertible preferred stock. Under the Redmile MRL, we agreed, among other things, to have Redmile’s restrictive legends on the shares of our capital stock removed. The foregoing rights and restrictions, as well as Redmile’s confidentiality obligations, will survive the closing of this offering. All other rights provided under the Redmile MRL will terminate upon the closing of this offering.
Our Relationship with Star Therapeutics LLC
We were formed as a wholly-owned subsidiary of Star Therapeutics LLC (Star LLC) in October 2018. In February 2022, we issued shares of our Series B redeemable convertible preferred stock to outside investors but remained a
180
Table of Contents
majority-owned subsidiary of Star LLC. In June 2023, Star LLC formed Electra LLC and contributed its shares of our Series A redeemable convertible preferred stock to Electra LLC in exchange for units of Electra LLC. Star LLC then distributed in-kind the units of Electra LLC to its members, certain of which are holders of more than 5% of our outstanding capital stock as described in the table below:
| HOLDERS |
PREFERRED UNITS OF ELECTRA LLC(1) |
|||
| OPI VII (2) |
6,813,384 | |||
| Redmile Biopharma Investments II, L.P. (2) |
6,752,194 | |||
| Westlake BioPartners Fund I, L.P. (3) |
12,002,003 | |||
| (1) | As of June 30, 2026, Electra LLC had 54,018,868 outstanding units. |
| (2) | Additional details regarding these stockholders and their equity holdings are provided in this prospectus under the section titled “Principal Stockholders” and above in the subsection titled “—Series C Redeemable Convertible Preferred Stock Financing.” |
| (3) | Westlake holds more than 5% of our outstanding capital stock. Dr. Seidenberg, a member of our board of directors, is the sole managing director of Westlake BioPartners GP II, LLC, the general partner of Westlake. Dr. Seidenberg has notified us that she intends to resign from our board of directors as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. |
As a result, we were spun out of and no longer owned by Star LLC. In January 2024, we entered into a Mutual Intercompany Services Agreement with Star Therapeutics, Inc. (Star Inc.), a wholly-owned operating entity of Star LLC (the ISA). The ISA replaced a prior intercompany services agreement between us and Star Inc. originally entered into in December 2018, as further amended in August 2021 and June 2023 (the Original Star Agreement, and together with the ISA, the Star Agreements). Pursuant to the Star Agreements, we agreed to provide certain services to Star Inc. and its affiliated entities, and Star Inc. agreed to provide certain services to us. During the period from January 1, 2023 through the termination of the ISA in October 2025, we paid an aggregate of approximately $15.5 million to Star Inc., and Star Inc. paid us an aggregate of approximately $0.2 million.
Employment Arrangements and Indemnification Agreements
We have entered into employment agreements and participation agreements to our Severance Plan with our executive officers. For more information regarding these agreements with our named executive officers, see the sections titled “Executive and Director Compensation—Employment Arrangements with Our Named Executive Officers,” and “Executive and Director Compensation—Potential Payments Upon Termination or Change of Control.”
Our amended and restated certificate of incorporation upon the closing of this offering will contain provisions limiting the liability of directors, and our amended and restated bylaws that will be in effect upon the closing of this offering will provide that we will indemnify each of our directors and officers to the fullest extent permitted under Delaware law. Our amended and restated certificate of incorporation and amended and restated bylaws upon the closing of this offering will also provide our board of directors with discretion to indemnify our employees and other agents when determined appropriate by the board.
In addition, we have entered, or intend to enter, into an indemnification agreement with each of our directors and executive officers, which requires us to indemnify them. For more information regarding these agreements, see the section titled “Executive and Director Compensation—Limitations of Liability and Indemnification Matters.”
Equity Awards to Directors and Executive Officers
We have granted equity awards to our directors and executive officers, as more fully described in the section titled “Executive and Director Compensation.”
Electra LLC has also granted equity awards through the grant of Profits Interest Units to our directors and executive officers. For a description of the 2023 Plan, see the section titled “Executive and Director Compensation – Electra Therapeutics LLC 2023 Equity Incentive Plan.” In addition, the following table summarizes the Profits Interest Units granted to our directors and executive officers.
181
Table of Contents
| PARTICIPANTS |
AGGREGATE PROFITS INTEREST UNITS |
MARKET VALUE ($) (6) |
||||||
| Quehuong (Kathy) Dong, Pharm.D., M.B.A. (1) |
2,345,871 | 2,426,291 | ||||||
| Gary S. Koe, Ph.D. (2) |
270,087 | 447,562 | ||||||
| Graham Parry, Ph.D. (3) |
486,156 | 369,478 | ||||||
| Adam Rosenthal, Ph.D. (4) |
671,828 | 886,813 | ||||||
| Nancy Stagliano, Ph.D. (5) |
1,080,348 | 1,805,424 | ||||||
| (1) | Dr. Dong is a member of our board of directors and our President and Chief Executive Officer. |
| (2) | Dr. Koe is our Chief Technical Officer. |
| (3) | Dr. Parry is our Chief Scientific Officer. |
| (4) | Dr. Rosenthal is a former member of our board of directors. In addition to Dr. Rosenthal’s Profits Interest Units, he also holds an aggregate of 5,000,000 fully vested Common Units of Electra LLC, which have a market value of $12,650,000, based on a fair market value of $2.53 per Common Unit as of December 31, 2025. |
| (5) | Dr. Stagliano is a member of our board of directors. |
| (6) | The market value is based on the fair market value of our Profits Interest Units as of December 31, 2025 of $2.53 per unit less the aggregate distribution threshold of the Profits Interest Units. |
Additionally, in November 2023, we entered into a letter agreement (the Rosenthal Letter Agreement) with Adam Rosenthal, Ph.D., a former member of our board of directors. Pursuant to the Rosenthal Letter Agreement, we granted Dr. Rosenthal the right to the continued vesting of certain unvested common units of Electra LLC previously granted to Dr. Rosenthal under the Electra LLC equity incentive plan, subject to his continued service as a member of our board of directors. In October 2025, we amended the Rosenthal Letter Agreement to provide that Dr. Rosenthal would also be eligible to receive an annual cash retainer of $25,000. The value of the continued vesting of the unvested common units of Electra LLC previously granted to Dr. Rosenthal under the Electra LLC equity incentive plan had a fair market value of $160,679.
LLC Distribution
Certain of our directors, executive officers and holders of more than 5% of our capital stock will directly or indirectly receive shares of our Series A redeemable convertible preferred stock in connection with the LLC Distribution. With respect to the distributions to Drs. Dong and Parry, each has agreed to enter into a Stock Restriction Agreement with us to subject Dr. Dong’s Unvested Shares and 7,106 shares Dr. Parry will receive in the LLC Distribution (Dr. Parry’s Unvested Shares), which reflects the number of shares of Series A redeemable convertible preferred stock that will be distributed to Dr. Parry with respect to his unvested Profits Interest Units of Electra LLC (assuming that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock), to vesting and a right of repurchase in our favor. For more information on the number of shares each of our directors, executive officers and holders of more than 5% of our capital stock will receive in connection with the LLC Distribution, see the section titled “Principal Stockholders,” and for a further discussion of Dr. Dong’s Unvested Shares, see the section titled “Executive and Director Compensation — Equity-based Incentive Awards.”
Policies and Procedures for Related Person Transactions
Prior to the closing of this offering, we will adopt a written related person transactions policy that sets forth our policies and procedures regarding the identification, review, consideration, and continuing oversight of “related person transactions.” For purposes of our policy only, a “related person transaction” is a transaction, arrangement, or relationship (or any series of similar transactions, arrangements, or relationships) in which we and any “related person” are participants involving an amount that exceeds the lesser of $120,000 or 1% of the average of our total assets at year-end for the last two completed fiscal years. Transactions involving compensation for services provided to us as an employee, consultant, or director are not considered related person transactions under this policy. A related person is any executive officer, director, nominee to become a director, or a holder of more than 5% of any class of our voting securities, including any of their immediate family members and affiliates, and entities owned or controlled by such persons or entities in which such person has a 5% or greater beneficial ownership interest.
182
Table of Contents
Under the policy, where a transaction has been identified as a related person transaction, management must present information regarding the proposed related person transaction to our audit committee (or, where review by our audit committee would be inappropriate for reasons of conflict of interest or otherwise, to another independent body of our board of directors) for review. The presentation must include a description of, among other things, all of the parties thereto, the direct and indirect interests of the related persons, the purpose of the transaction, the material facts, the benefits of the transaction to us and whether any alternative transactions are available, an assessment of whether the terms are comparable to the terms available from unrelated third parties or to employees generally, and management’s recommendation. To identify related person transactions in advance, we rely on information supplied by our executive officers, directors, and certain significant stockholders. In considering related person transactions, our audit committee or another independent body of our board of directors takes into account the relevant available facts and circumstances including, but not limited to:
| ∎ | the risks, costs, and benefits to us; |
| ∎ | the impact on a director’s independence in the event the related person is a director, immediate family member of a director, or an entity with which a director is affiliated; |
| ∎ | the terms of the transaction; |
| ∎ | the availability of other sources for comparable services or products; and |
| ∎ | the terms available to or from, as the case may be, unrelated third parties. |
In the event a director has an interest in the proposed transaction, the director must recuse himself or herself from the deliberations and approval.
183
Table of Contents
The following table sets forth information with respect to the beneficial ownership of our capital stock as of August 1, 2026, as adjusted to reflect the sale of our common stock offered by us in this offering assuming no exercise of the underwriters’ option to purchase additional shares, for:
| ∎ | each of our named executive officers; |
| ∎ | each of our current directors; |
| ∎ | all of our current executive officers and directors as a group; and |
| ∎ | each person or group of affiliated persons known by us to beneficially own more than 5% of our common stock. |
We have determined beneficial ownership in accordance with the rules and regulations of the SEC, and the information is not necessarily indicative of beneficial ownership for any other purpose. Except as indicated by the footnotes below, we believe, based on information furnished to us, that the persons and entities named in the table below have sole voting and sole investment power with respect to all shares that they beneficially own, subject to applicable community property laws.
Applicable percentage ownership before the offering is based on 39,460,916 shares of our common stock outstanding as of August 1, 2026, assuming the automatic conversion of all outstanding shares of our redeemable convertible preferred stock into 39,404,292 shares of our common stock immediately prior to the closing of this offering. Applicable percentage ownership after the offering is based on the sale of 21,666,667 shares of our common stock in this offering, assuming no exercise by the underwriters of their option to purchase additional shares. In computing the number of shares beneficially owned by a person and the percentage ownership of such person, we deemed to be outstanding all shares subject to options held by the person that are currently exercisable, or exercisable within 60 days of August 1, 2026. However, except as described above, we did not deem such shares outstanding for the purpose of computing the percentage ownership of any other person.
Unless otherwise indicated, the address of each beneficial owner listed below is c/o Electra Therapeutics, Inc., 230 E Grand Avenue, Suite S-100, South San Francisco, California 94080. We believe, based on information provided to us, that each of the stockholders listed below has sole voting and investment power with respect to the shares beneficially owned by the stockholder unless noted otherwise, subject to community property laws where applicable.
The following table does not reflect any potential purchases by our executive officers, directors, their affiliated entities, holders of more than 5% of our common stock in this offering, or any equity awards granted to our executive officers or directors contingent on this offering. If any shares are purchased by and to the extent any such equity awards have been granted to these persons or entities, the number and percentage of shares of our common stock beneficially owned by them after this offering will differ from the amounts set forth in the following table.
| SHARES BENEFICIALLY OWNED PRIOR TO OFFERING |
SHARES BENEFICIALLY OWNED AFTER OFFERING |
|||||||||||||||
| NAME OF BENEFICIAL OWNER |
NUMBER | PERCENTAGE | NUMBER | PERCENTAGE | ||||||||||||
| 5% or Greater Stockholders: |
||||||||||||||||
| Electra Therapeutics LLC (1) |
15,065,741 | 38.2 | % | — | — | |||||||||||
| Entities affiliated with OrbiMed (2) |
3,549,527 | 9.0 | % | 5,548,593 | 9.1 | % | ||||||||||
| Entities affiliated with Westlake BioPartners Fund I, L.P. (3) |
2,820,403 | 7.1 | % | 6,341,824 | 10.4 | % | ||||||||||
| Redmile Biopharma Investments II, L.P. (4) |
2,811,887 | 7.1 | % | 4,793,000 | 7.8 | % | ||||||||||
| Aventis Inc. (5) |
2,461,948 | 6.2 | % | 2,461,948 | 4.0 | % | ||||||||||
| LSP 7 Coöperatief U.A. (6) |
2,461,948 | 6.2 | % | 2,461,948 | 4.0 | % | ||||||||||
| Entities affiliated with New Leaf Ventures (7) |
1,722,233 | 4.4 | % | 3,245,831 | 5.3 | % | ||||||||||
184
Table of Contents
| SHARES BENEFICIALLY OWNED PRIOR TO OFFERING |
SHARES BENEFICIALLY OWNED AFTER OFFERING |
|||||||||||||||
| NAME OF BENEFICIAL OWNER |
NUMBER | PERCENTAGE | NUMBER | PERCENTAGE | ||||||||||||
| Named Executive Officers and Directors: |
||||||||||||||||
| Quehuong (Kathy) Dong, Pharm.D., M.B.A. (8) |
1,399,175 | 3.4 | % | 1,853,543 | 3.0 | % | ||||||||||
| Gary S. Koe, Ph.D. (9) |
380,837 | * | 444,364 | * | ||||||||||||
| Kim-Hien Dao, D.O., Ph.D. (10) |
547,202 | 1.4 | % | 547,202 | * | |||||||||||
| Matthew Fust, M.B.A. (11) |
136,800 | * | 136,800 | * | ||||||||||||
| Thomas Geninatti, Ph.D.(12) |
— | — | — | — | ||||||||||||
| Carl L. Gordon, Ph.D., C.F.A. (13) |
3,549,527 | 9.0 | % | 5,548,593 | 9.1 | % | ||||||||||
| Beth Seidenberg, M.D. (14) |
2,820,403 | 7.1 | % | 6,341,824 | 10.4 | % | ||||||||||
| Nancy Stagliano, Ph.D. (15) |
383,352 | * | 638,472 | 1.0 | % | |||||||||||
| All directors and executive officers as a group (10 persons) (16) |
9,994,008 | 23.2 | % | 16,372,783 | 25.0 | % | ||||||||||
| * | Represents beneficial ownership of less than 1%. |
| (1) | Consists of 15,065,741 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock. Prior to the closing of this offering, Electra LLC expects to distribute shares of Series A redeemable convertible preferred stock to its members (the LLC Distribution). Each of OPI VII, Westlake BioPartners Fund I, LP (Westlake Fund I), Redmile, New Leaf Biopharma Opportunities II, L.P. (New Leaf II), New Leaf Ventures IV, L.P. (New Leaf IV), Dr. Stagliano, Dr. Dong, Dr. Koe, and certain other executive officers hold outstanding units in Electra LLC and will receive shares of Series A redeemable convertible preferred stock in connection with the LLC Distribution, in each case, as further described in their respective footnotes below. Electra LLC is governed by a board of managers comprised of Dr. Dong, Dr. Gordon, Dr. Seidenberg, and Dr. Stagliano. No individual board member holds sole or shared voting or dispositive control over the shares held by Electra LLC, and each expressly disclaims beneficial ownership of such shares. |
| (2) | The number of shares beneficially owned prior to the offering consists of (i) 1,087,934 shares of common stock issuable upon conversion of our Series B redeemable convertible preferred stock held by OPI VII, (ii) 1,703,314 shares of common stock issuable upon conversion of our Series C redeemable convertible preferred stock held by OPI VII, and (iii) 758,279 shares of common stock issuable upon conversion of our Series C redeemable convertible preferred stock held by Orbi Genesis, but excludes any shares to be distributed to OPI VII in connection with the LLC Distribution. In connection with the LLC Distribution, OPI VII expects to receive 1,999,066 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number of shares beneficially owned after the offering. GP VII is the general partner of OPI VII. Genesis GP is the general partner of Orbi Genesis and OrbiMed Advisors is the managing member of GP VII and Genesis GP. By virtue of such relationships, GP VII, Genesis GP, and OrbiMed Advisors may be deemed to have voting power and investment power over the shares held by OPI VII and Orbi Genesis, and may be deemed to have voting power and investment power over the shares held by OPI VII following the LLC Distribution, and as a result, may be deemed to have beneficial ownership of such shares. OrbiMed Advisors exercises voting and investment power through a management committee comprised of Carl L. Gordon, Sven H. Borho, and W. Carter Neild, each of whom disclaims beneficial ownership of the shares held by OPI VII and Orbi Genesis. The principal business address of each of the foregoing entities and individuals is 601 Lexington Avenue, 54th Floor, New York, NY 10022. |
| (3) | The number of shares beneficially owned prior to the offering consists of (i) 1,624,880 shares of common stock issuable upon conversion of our Series B redeemable convertible preferred stock and (ii) 1,195,523 shares of common stock issuable upon conversion of our Series C redeemable convertible preferred stock held by Westlake BioPartners Fund II, LP (Westlake Fund II), but excludes any shares to be distributed to Westlake Fund I in connection with the LLC Distribution. In connection with the LLC Distribution, Westlake Fund I expects to receive 3,521,421 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number of shares beneficially owned after the offering. Westlake BioPartners GP II, LLC (Westlake GP II), the general partner of Westlake Fund II, may be deemed to have sole voting and dispositive power over the shares held by Westlake Fund II. Westlake BioPartners GP I, LLC (Westlake GP I), the general partner of Westlake Fund I, may be deemed to have sole voting and dispositive power over the shares held by Westlake Fund I following the LLC Distribution. Dr. Seidenberg, the sole managing director of Westlake GP II and Westlake GP I, may be deemed to have the power to vote and dispose of these shares. The principal business address for each of the foregoing entities and individuals is c/o Westlake BioPartners, LLC, 3075 Townsgate Road, Suite 140, Westlake Village, California 91361. |
| (4) | The number of shares beneficially owned prior to the offering consists of (i) 1,107,872 shares of common stock issuable upon conversion of our Series B redeemable convertible preferred stock and (ii) 1,704,015 shares of common stock issuable upon conversion of our Series C redeemable convertible preferred stock held by Redmile, but excludes any shares to be distributed to Redmile in connection with the LLC Distribution. In connection with the LLC Distribution, Redmile expects to receive 1,981,113 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number of shares beneficially owned after the offering. The shares held by Redmile may be deemed beneficially owned by Redmile Group, LLC (Redmile Group) as |
185
Table of Contents
| investment manager of Redmile, and may be deemed to beneficially own the shares held by Redmile following the LLC Distribution. The reported securities may also be deemed beneficially owned by Jeremy Green as the principal of Redmile Group, who may also be deemed to beneficially own the shares held by Redmile following the LLC Distribution. Each of Redmile Group and Mr. Green disclaims beneficial ownership of the reported securities except to the extent of their pecuniary interest therein. The principal business address for each of the foregoing entities and individuals is 900 Larkspur Landing Circle, Suite 270, Larkspur, CA 94939. |
| (5) | Consists of 2,461,948 shares of common stock issuable upon conversion of our Series C redeemable convertible preferred stock. Sanofi is a wholly-owned subsidiary of Sanofi S.A., a French société anonyme (limited liability company), which may be deemed to have voting and dispositive power over the shares held by Sanofi. The address of Sanofi is c/o Sanofi—Global Alliance Management, 450 Water Street, Cambridge, MA 02141. |
| (6) | Consists of 2,461,948 shares of common stock issuable upon conversion of our Series C redeemable convertible preferred stock. As the sole director of LSP 7 Coöperatief UA, LSP 7 Management BV may be deemed to beneficially own these shares. As managing directors of LSP 7 Management BV, each of Merijn Klaassen, René Kuijten, and Joachim Rothe may also be deemed to beneficially own these shares. Each of Merijn Klaassen, René Kuijten, and Joachim Rothe disclaims beneficial ownership of the reported securities. The business address of LSP 7 Coöperatief UA is Johannes Vermeerplein 9 1071 DV Amsterdam, Netherlands. |
| (7) | The number of shares beneficially owned prior to the offerings consists of (a) (i) 203,109 shares of common stock issuable upon the conversion of our Series B redeemable convertible preferred stock and (ii) 1,367,536 shares of common stock issuable upon conversion of our Series C redeemable convertible preferred stock held by New Leaf II, and (b) (i) 110,787 shares of common stock issuable upon conversion of our Series B redeemable convertible preferred stock and (ii) 40,801 shares of common stock issuable upon conversion of Series C redeemable convertible preferred stock held by New Leaf IV, but, in each case, excludes any shares to be distributed to New Leaf II and New Leaf IV in connection with the LLC Distribution. In connection with the LLC Distribution, (x) New Leaf II expects to receive 237,733 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock and (y) New Leaf IV expects to receive 1,285,865 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, each of which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number shares beneficially owned after the offering. |
| (8) | The number of shares beneficially owned prior to the offering consists of 1,399,175 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026, but excludes any shares to be distributed to Dr. Dong in connection with the LLC Distribution. In connection with the LLC Distribution, Dr. Dong expects to receive 454,368 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, of which 72,949 shares of Series A redeemable convertible preferred stock remain subject to a right of repurchase in our favor, and which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number of shares beneficially owned after the offering. |
| (9) | The number of shares beneficially owned prior to the offering consists of 380,837 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026, but excludes any shares to be distributed to Dr. Koe in connection with the LLC Distribution. In connection with the LLC Distribution, Dr. Koe expects to receive 63,527 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number of shares beneficially owned after the offering. |
| (10) | Consists of 547,202 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026. |
| (11) | Consists of 136,800 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026. |
| (12) | Dr. Geninatti has notified us that he intends to resign from our board of directors effective as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. |
| (13) | Consists of the shares described in footnote (2) above. |
| (14) | Consists of the shares described in footnote (3) above. Dr. Seidenberg has notified us that she intends to resign from our board of directors effective as of immediately prior to the effectiveness of the registration statement of which this prospectus forms a part. |
| (15) | Consists of (i) 288,814 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026 held by Dr. Stagliano and (ii) 94,538 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026 held by The Nancy E. Stagliano Trust, but excludes any shares to be distributed to Dr. Stagliano in connection with the LLC Distribution. Dr. Stagliano is the trustee of The Nancy E. Stagliano Trust. In connection with the LLC Distribution, Dr. Stagliano expects to receive 255,120 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number of shares beneficially owned after the offering. |
| (16) | Consists of the shares described in footnotes (8)-(15) above as well as (i) 456,003 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026, held by Chris Clark and (ii) 320,709 shares of common stock subject to options that are exercisable within 60 days of August 1, 2026 held by Graham Parry, Ph.D., but excludes any shares to be distributed to certain of our officers in connection with the LLC Distribution. In addition to the shares of common stock described in footnotes (8)-(15), in connection with the LLC Distribution, Dr. Parry expects to receive 85,273 shares of common stock issuable upon conversion of our Series A redeemable convertible preferred stock, of which 7,106 shares of Series A redeemable convertible preferred stock remain subject to a right of repurchase in our favor, and which assumes that Electra LLC uses the midpoint of the range set forth on the cover of this prospectus for purposes of valuing a share of Series A redeemable convertible preferred stock. The shares received upon the LLC Distribution are included in the number of shares beneficially owned after the offering. |
186
Table of Contents
Upon the filing of our amended and restated certificate of incorporation and the closing of this offering, our authorized capital stock will consist of 900,000,000 shares of our common stock, par value $0.0001 per share, and 10,000,000 shares of preferred stock, par value $0.0001 per share. All of our authorized preferred stock upon the closing of this offering will be undesignated. The following is a summary of the rights of our common and preferred stockholders and some of the provisions of our amended and restated certificate of incorporation and amended and restated bylaws, which will become effective upon the closing of this offering, and of the DGCL. This summary is not complete. For more detailed information, please see our amended and restated certificate of incorporation and amended and restated bylaws, which are filed as exhibits to the registration statement of which this prospectus is a part, as well as the relevant provisions of the DGCL.
Common Stock
Outstanding Shares
As of June 30, 2026, we had 29,329 shares of our common stock outstanding held of record by stockholders. This amount excludes our outstanding shares of redeemable convertible preferred stock, which will automatically convert into shares of our common stock immediately prior to the closing of this offering. Based on the number of shares of our common stock outstanding as of June 30, 2026, and assuming (i) the automatic conversion of all of our outstanding shares of redeemable convertible preferred stock, (ii) the issuance by us of 21,666,667 shares of our common stock in this offering, and (iii) no exercise by the underwriters of their option to purchase additional shares, there will be 61,100,288 shares of our common stock outstanding upon the closing of this offering.
Voting
Each holder of our common stock is entitled to one vote for each share on all matters submitted to a vote of stockholders, except as otherwise expressly provided in our amended and restated certificate of incorporation or required by applicable law. There are no cumulative voting rights.
Dividends
Subject to preferences that may be applicable to any then-outstanding preferred stock, the holders of our common stock are entitled to receive dividends, if any, as may be declared from time to time by our board of directors out of legally available funds.
Liquidation
In the event of our liquidation, dissolution, or winding-up, holders of our common stock will be entitled to share ratably in the net assets legally available for distribution to stockholders after the payment of all of our debts and other liabilities, subject to the satisfaction of any liquidation preference granted to the holders of any outstanding shares of preferred stock.
Rights, Preferences, and Privileges
Holders of our common stock have no preemptive, conversion, or subscription rights, and there are no redemption or sinking fund provisions applicable to our common stock. The rights, preferences, and privileges of the holders of our common stock are subject to, and may be adversely affected by, the rights of the holders of shares of any series of our preferred stock that we may designate and issue in the future.
Fully Paid and Nonassessable
All of our outstanding shares of our common stock are, and the shares of our common stock to be issued in this offering will be, fully paid and nonassessable.
Redeemable Convertible Preferred Stock
As of June 30, 2026, there were 39,404,292 shares of redeemable convertible preferred stock outstanding, held of record by 23 stockholders.
Immediately prior to the closing of this offering, all outstanding shares of redeemable convertible preferred stock will automatically convert into 39,404,292 shares of our common stock.
187
Table of Contents
Under the amended and restated certificate of incorporation that will become effective upon the closing of this offering, our board of directors will have the authority, without further action by the stockholders, to issue up to 10,000,000 shares of preferred stock in one or more series, to establish from time to time the number of shares to be included in each such series, to fix the rights, preferences, and privileges of the shares of each wholly unissued series and any qualifications, limitations, or restrictions thereon, and to increase or decrease the number of shares of any such series, but not below the number of shares of such series then outstanding.
Our board of directors may authorize the issuance of preferred stock with voting or conversion rights that could adversely affect the voting power or other rights of the holders of the common stock. The issuance of preferred stock, while providing flexibility in connection with possible acquisitions and other corporate purposes, could, among other things, have the effect of delaying, deferring, or preventing a change in our control that may otherwise benefit holders of our common stock and may adversely affect the market price of the common stock and the voting and other rights of the holders of our common stock. We have no current plans to issue any shares of preferred stock.
Stock Options
As of June 30, 2026, 5,701,943 shares of our common stock were issuable upon the exercise of outstanding stock options, issued under our 2022 Plan, at a weighted-average exercise price of $4.92 per share and 259,921 shares of our common stock were issuable upon the exercise of stock options outstanding granted under the 2022 Plan subsequent to June 30, 2026, with a weighted-average exercise price of $7.03 per share. For information regarding the terms of our equity incentive plans, see the section titled “Executive and Director Compensation—Equity Benefit Plans.”
Registration Rights
Upon the closing of this offering, certain holders of 39,404,292 shares of our common stock, including those issuable upon the automatic conversion of our redeemable convertible preferred stock, will be entitled to rights with respect to the registration of these securities under the Securities Act. These rights are provided under the terms of an amended and restated investors’ rights agreement (investor rights agreement) between us and holders of our redeemable convertible preferred stock. These shares are collectively herein referred to as registrable securities. The investor rights agreement includes demand registration rights, Form S-3 registration rights, and piggyback registration rights. All fees, costs, and expenses of underwritten registrations under this agreement will be borne by us and all selling expenses, including underwriting discounts and selling commissions, will be borne by the holders of the shares being registered.
Form S-1 Demand Registration Rights
Beginning six months after the effective date of the registration statement of which this prospectus is a part, certain holders of registrable securities are entitled to demand registration rights. Under the terms of the investor rights agreement, we will be required, upon the request of the holders of a majority of our outstanding registrable securities, as defined in the investor rights agreement, to file a registration statement with respect to outstanding registrable securities having an anticipated aggregate gross offering price to the public of at least $15 million, and use commercially reasonable efforts to effect, as soon as practicable, the registration of all or a portion of their registrable securities for public resale. We are not obligated to take any action in response to such request (i) during the period starting from 60 days prior to our good faith estimate of the date of filing of, and ending 180 days after the effective date of, our initial public offering, (ii) if we have already effected two registrations pursuant to such requests for registration on Form S-1, or (iii) if the initiating holders propose to dispose of shares of registrable securities that may be immediately registered on Form S-3. Additionally, if we furnish to the holders requesting such registration a certificate signed by our Chief Executive Officer or Chairperson stating that, in the good faith judgment of our board of directors, it would be seriously detrimental to us and our stockholders for such registration statement to either become effective at such time, then we shall have the right to defer taking action with respect to such filing, and any time periods with respect to filing or effectiveness thereof shall be tolled correspondingly, for a period of not more than 120 days after the request of the initiating holders is given; provided, however, that we may not invoke this right more than once in any 12-month period, as described in the investor rights agreement.
Form S-3 Demand Registration Rights
Pursuant to the investor rights agreement, if we are eligible to file a registration statement on Form S-3, we will be required, upon request of the holders of at least 20% of our outstanding registrable securities, as defined in the
188
Table of Contents
investor rights agreement, to register their registrable securities under the Securities Act so long as the total amount of registrable shares requested to be registered has an anticipated aggregate offering price to the public, net of any underwriters’ discounts or commissions, of at least $10 million. We are not obligated to take any action in response to such request (i) during the period starting from 30 days prior to our good faith estimate of the date of the filing of and ending on a date 90 days following the effective date of, a registration initiated by us; (ii) if we have, within the 12-month period preceding the date of such request, already effected two registrations pursuant to such requests for registration on Form S-3; and (iii) if, within 30 days of receipt of a written request from initiating holders, we give notice to such holders of our intention to make a public offering within 120 days. Additionally, if we furnish to the holders requesting such registration a certificate signed by our Chief Executive Officer or Chairperson stating that, in the good faith judgment of our board of directors, it would be seriously detrimental to us and our stockholders for such registration statement to become effective at such time, then we shall have the right to defer such filing, for a period of not more than 90 days after the request of the initiating holders is given; provided, however, that we may not invoke this right more than once in any 12-month period, as described in the investor rights agreement; and provided further that we shall not register any securities for our own account or any other stockholder during such 90 day period.
Piggyback Registration Rights
If we propose to register any of our securities under the Securities Act in another offering solely for cash, either for our own account or for the account of other security holders, the holders of registrable securities will be entitled to notice of the registration and will be entitled to include their shares of our common stock in the registration, subject to certain conditions and limitations, including the right of the underwriters to limit the number of shares included in such registration under specified circumstances.
Anti-Takeover Effects of Provisions of our Amended and Restated Certificate of Incorporation, our Amended and Restated Bylaws, and Delaware Law
Delaware Anti-Takeover Law
We are subject to Section 203 of the DGCL (Section 203). Section 203 generally prohibits a public Delaware corporation from engaging in a “business combination” with an “interested stockholder” for a period of three years following the time that such stockholder became an interested stockholder, unless:
| ∎ | prior to such time the board of directors of the corporation approved either the business combination or the transaction which resulted in the stockholder becoming an interested stockholder; |
| ∎ | upon consummation of the transaction which resulted in the stockholder becoming an interested stockholder, the interested stockholder owned at least 85% of the voting stock of the corporation outstanding at the time the transaction commenced, excluding for purposes of determining the voting stock outstanding (but not the outstanding voting stock owned by the interested stockholder) those shares owned (i) by persons who are directors and also officers and (ii) employee stock plans in which employee participants do not have the right to determine confidentially whether shares held subject to the plan will be tendered in a tender or exchange offer; or |
| ∎ | at or subsequent to such time, the business combination is approved by the board of directors and authorized at an annual or special meeting of stockholders, and not by written consent, by the affirmative vote of at least 66-2/3% of the outstanding voting stock which is not owned by the interested stockholder. |
Section 203 defines a “business combination” to include (subject to exceptions):
| ∎ | any merger or consolidation involving the corporation or any direct or indirect majority-owned subsidiary of the corporation and the interested stockholder; |
| ∎ | any sale, transfer, pledge, or other disposition of 10% or more of the assets of the corporation involving the interested stockholder (in one transaction or a series of transactions); |
| ∎ | any transaction that results in the issuance or transfer by the corporation or by any direct or indirect majority-owned subsidiary of the corporation of any stock of the corporation or of such subsidiary to the interested stockholder; |
189
Table of Contents
| ∎ | any transaction involving the corporation or any direct or indirect majority-owned subsidiary of the corporation that has the effect, directly or indirectly, of increasing the proportionate share of the stock of any class or series of the corporation beneficially owned by the interested stockholder; and |
| ∎ | the receipt by the interested stockholder of the benefit, directly or indirectly, of any loans, advances, guarantees, pledges, or other financial benefits provided by or through the corporation. |
In general, Section 203 defines an “interested stockholder” as any entity or person who, together with the person’s affiliates and associates, beneficially owns, or within the three years prior to the time of determination of interested stockholder status did own, 15% or more of the outstanding voting stock of the corporation.
A Delaware corporation may “opt out” of these provisions with an express provision in its original certificate of incorporation or an express provision in its amended and restated certificate of incorporation or amended and restated bylaws resulting from a stockholders’ amendment approved by at least a majority of the outstanding voting shares. We have not opted out of these provisions. As a result, mergers or other takeover or change in control attempts of us may be discouraged or prevented.
Amended and Restated Certificate of Incorporation and Amended and Restated Bylaws
Provisions of our amended and restated certificate of incorporation and amended and restated bylaws, which will become effective upon the closing of this offering, may delay or discourage transactions involving an actual or potential change in our control or change in our management, including transactions in which stockholders might otherwise receive a premium for their shares or transactions that our stockholders might otherwise deem to be in their best interests. Therefore, these provisions could adversely affect the price of our common stock. Among other things, our amended and restated certificate of incorporation and amended and restated bylaws:
| ∎ | permit our board of directors to issue up to 10,000,000 shares of preferred stock, with any rights, preferences, and privileges as they may designate (including the right to approve an acquisition or other change in our control); |
| ∎ | provide that the authorized number of directors may be changed only by resolution of the board of directors; |
| ∎ | provide that the board of directors or any individual director may only be removed with cause and the affirmative vote of the holders of at least 66-2/3% of the voting power of all of our then outstanding common stock; |
| ∎ | provide that all vacancies, including newly created directorships, may, except as otherwise required by law, be filled by the affirmative vote of a majority of directors then in office, even if less than a quorum; |
| ∎ | divide our board of directors into three classes with each class serving three-year staggered terms; |
| ∎ | require that any action to be taken by our stockholders must be effected at a duly called annual or special meeting of stockholders and not be taken by written consent; |
| ∎ | provide that stockholders seeking to present proposals before a meeting of stockholders or to nominate candidates for election as directors at a meeting of stockholders must provide notice in writing in a timely manner and also specify requirements as to the form and content of a stockholder’s notice; |
| ∎ | do not provide for cumulative voting rights (therefore allowing the holders of a majority of the shares of our common stock entitled to vote in any election of directors to elect all of the directors standing for election, if they should so choose); |
| ∎ | provide that special meetings of our stockholders may be called only by the Chairperson of the board, our Chief Executive Officer or by the board of directors pursuant to a resolution adopted by a majority of the total number of authorized directors; |
| ∎ | provide that, unless we consent in writing to the selection of an alternative forum, the Court of Chancery of the State of Delaware and any appellate court therefrom shall be the sole and exclusive forum for: (i) any derivative claim or cause of action brought on our behalf; (ii) any claim or cause of action that is based upon a violation of a duty owed by any of our current or former directors, officers, employees, or stockholders, to us or our stockholders; (iii) any claim or cause of action against us or any of our current or former directors, officers, or other employees, arising out of or pursuant to any provision of the DGCL, our amended and restated certificate of incorporation, or our bylaws; (iv) any claim or cause of action seeking to interpret, apply, enforce, or determine the validity of our amended and restated certificate of incorporation |
190
Table of Contents
| or our bylaws; (v) any claim or cause of action as to which the DGCL confers jurisdiction to the Court of Chancery of the State of Delaware; and (vi) any claim or cause of action or proceeding asserting a claim against us or any of our current or former directors, officers, or other employees governed by the internal-affairs doctrine or otherwise related to our internal affairs, in all cases to the fullest extent permitted by applicable law and subject to the court’s having personal jurisdiction over the indispensable parties named as defendants; provided, however, that if the designation of such court as the sole and exclusive forum for a claim or action referred to in foregoing clauses (i) through (vi) would violate applicable law, then the U.S. District Court for the District of Delaware shall be the sole and exclusive forum for such claim or cause of action; provided further, that the exclusive forum provision does not apply to suits brought to enforce a duty or liability created by the Exchange Act; and |
| ∎ | provide that unless we consent in writing to the selection of an alternative forum, to the fullest extent permitted by law, the federal district courts of the United States of America shall be the exclusive forum for the resolution of any complaint asserting a cause of action arising under the Securities Act, including all causes of action asserted against any defendant named in such complaint. For the avoidance of doubt, this provision is intended to benefit and may be enforced by us, our officers and directors, the underwriters for any offering giving rise to such complaint, and any other professional entity whose profession gives authority to a statement made by that person or entity and who has prepared or certified any part of the documents underlying the offering. |
The amendment of any of these provisions, with the exception of the ability of our board of directors to issue shares of preferred stock and designate any rights, preferences and privileges thereto, would require approval by the holders of at least 66-2/3% of our then-outstanding common stock.
Exchange Listing
We have applied to list our common stock on the Nasdaq Global Select Market under the symbol “ETRA.” We believe that upon the closing of this offering, we will meet the standards for listing on Nasdaq, and the closing of this offering is contingent upon such listing.
Transfer Agent and Registrar
Upon the closing of this offering, the transfer agent and registrar for our common stock will be Computershare Trust Company, N.A.. The transfer agent and registrar’s address is 150 Royall Street, Canton, Massachusetts 02021, and its telephone number is (781) 575-4019.
191
Table of Contents
SHARES ELIGIBLE FOR FUTURE SALE
Prior to this offering, there has been no public market for our common stock. Future sales of substantial amounts of our common stock in the public market, or the perception that such sales may occur, could adversely affect the market price of our common stock. Furthermore, because only a limited number of shares of our common stock will be available for sale shortly after this offering due to certain contractual and legal restrictions on resale described below, sales of substantial amounts of our common stock in the public market after such restrictions lapse, or the anticipation of such sales, could adversely affect the prevailing market price of our common stock and our ability to raise equity capital in the future. Although we have applied to have our common stock listed on the Nasdaq Global Select Market, we cannot assure you that there will be an active public market for our common stock.
Following the closing of this offering, based on the number of shares of our common stock outstanding as of June 30, 2026 and assuming (i) the issuance of 21,666,667 shares of our common stock in this offering, (ii) the automatic conversion of all outstanding shares of our redeemable convertible preferred stock into 39,404,292 shares of our common stock, which will automatically occur immediately prior to the closing of the offering, and (iii) no exercise of the underwriters’ option to purchase additional shares, we will have an aggregate of 61,100,288 shares of our common stock outstanding.
Of these shares, all shares of our common stock sold in this offering will be freely tradable without restriction or further registration under the Securities Act, except for any shares of our common stock purchased by our “affiliates,” as that term is defined in Rule 144 under the Securities Act or any shares, subject to lock-up agreements.
Shares purchased by our affiliates would be subject to the Rule 144 resale restrictions described below, other than the holding period requirement.
The remaining shares of our common stock outstanding after this offering will be “restricted securities,” as that term is defined in Rule 144 under the Securities Act. These restricted securities are eligible for public sale only if they are registered under the Securities Act or if they qualify for an exemption from registration under Rule 144 or Rule 701 under the Securities Act, each of which is summarized below and, if subject to lock-up agreements, may only be sold after the expiration of the 180-day lock-up period. We expect that substantially all of these shares will be subject to a 180-day lock-up period under the lock-up and market stand-off agreements described below.
We may issue shares of our common stock from time to time as consideration for future acquisitions, investments, or other corporate purposes. In the event any such acquisition, investment, or other transaction is significant, the number of shares of our common stock that we may issue may also be significant. We may also grant registration rights covering those shares of our common stock issued in connection with any such acquisition, investment, or other transaction.
In addition, shares of our common stock that are either subject to outstanding options or reserved for future issuance under our equity incentive plans will become eligible for sale in the public market to the extent permitted by the provisions of various vesting schedules, the lock-up agreements described below and Rules 144 and 701 under the Securities Act.
Rule 144
In general, under Rule 144 as currently in effect, once we have been subject to public company reporting requirements of Section 13 or Section 15(d) of the Exchange Act for at least 90 days, an eligible stockholder is entitled to sell such shares without complying with the manner of sale, volume limitation or notice provisions of Rule 144, subject to compliance with the public information requirements of Rule 144. To be an eligible stockholder under Rule 144, such stockholder must not be deemed to have been one of our affiliates for purposes of the Securities Act at any time during the 90 days preceding a sale and must have beneficially owned the shares proposed to be sold for at least six months, including the holding period of any prior owner other than our affiliates. If such a person has beneficially owned the shares proposed to be sold for at least one year, including the holding period of any prior owner other than our affiliates, then such person is entitled to sell such shares without complying with any of the requirements of Rule 144, subject to the expiration of the lock-up agreements described below.
192
Table of Contents
In general, under Rule 144, as currently in effect, our affiliates or persons selling shares on behalf of our affiliates will be entitled to sell shares on expiration of the lock-up agreements described below. Beginning 90 days after the date of this prospectus, within any three-month period, such stockholders may sell a number of shares that does not exceed the greater of:
| ∎ | 1% of the number of shares of our common stock then outstanding, which will equal approximately 611,003 shares immediately after this offering; or |
| ∎ | the average weekly trading volume in our common stock on the Nasdaq Global Select Market during the four calendar weeks preceding the filing of a notice on Form 144 with respect to such sale, provided in each case that we have been subject to the Exchange Act periodic reporting requirements for at least 90 days before the sale. |
Sales under Rule 144 by our affiliates or persons selling shares on behalf of our affiliates are also subject to certain manner of sale provisions and notice requirements and to the availability of current public information about us.
Rule 701
Rule 701 generally allows a stockholder who was issued shares under a written compensatory plan or contract, and who is not deemed to have been an affiliate of our company during the immediately preceding 90 days, to sell these shares in reliance on Rule 144, but without being required to comply with the public information, holding period, volume limitation, or notice provisions of Rule 144. Rule 701 also permits our affiliates to sell their Rule 701 shares under Rule 144 without complying with the holding period requirements of Rule 144. All holders of Rule 701 shares, however, are required by that rule to wait until 90 days after the date of this prospectus before selling those shares under Rule 701, subject to the expiration of the lock-up agreements described below and in the section titled “Underwriting.”
Form S-8 Registration Statement
We intend to file one or more registration statements on Form S-8 under the Securities Act to register all shares of our common stock subject to outstanding stock options and common stock issued or issuable under the 2022 Plan, the 2026 Plan, and the ESPP. We expect to file the registration statement covering shares offered pursuant to these stock plans shortly after the date of this prospectus, permitting the resale of such shares by non-affiliates in the public market without restriction under the Securities Act and the sale by affiliates in the public market subject to compliance with the resale provisions of Rule 144.
Lock-Up Agreements
We, our directors, executive officers, and the holders of substantially all of our equity securities, have agreed with the underwriters that for a period of 180 days after the date of this prospectus, subject to specified exceptions as detailed further in the section titled “Underwriting,” we will not, except with the prior written consent of Jefferies LLC, TD Securities (USA) LLC and Evercore Group L.L.C., and they will not, except with the prior written consent of Jefferies LLC and TD Securities (USA) LLC, offer, pledge, sell, contract to sell, sell any option or contract to purchase, purchase any option or contract to sell, grant any option, right, or warrant to sell or otherwise dispose of, or transfer any shares of our common stock or any securities convertible into or exercisable or exchangeable for shares of our common stock, request or demand that we file a registration statement related to our common stock, or enter into any swap or other agreement that transfers to another, in whole or in part, directly or indirectly, the economic consequence of ownership of the common stock. Substantially all of our optionholders are subject to a market stand-off agreement with us, which imposes similar restrictions.
Upon expiration of the lock-up period, certain of our stockholders will have the right to require us to register their shares under the Securities Act. See the sections titled “Registration Rights” below and “Description of Capital Stock—Registration rights.”
Upon the expiration of the lock-up period, substantially all of the shares subject to such lock-up restrictions will become eligible for sale, subject to the limitations discussed above.
193
Table of Contents
Registration Rights
Upon the closing of this offering and the expiration or release from the terms of applicable lock-up agreements, holders of an aggregate of 39,404,292 shares of our common stock, which includes all of the shares of our common stock issuable upon the automatic conversion of our redeemable convertible preferred stock immediately prior to the closing of this offering, or their transferees, will be entitled to various rights with respect to the registration of these shares under the Securities Act. Registration of these shares under the Securities Act would result in these shares becoming fully tradable without restriction under the Securities Act immediately upon the effectiveness of the registration, except for shares subsequently purchased by affiliates. See the section titled “Description of Capital Stock—Registration Rights” for additional information.
Rule 10b5-1 Sales Plans
After the closing of this offering, certain of our employees, including our executive officers, and/or directors, may enter into written trading plans that are intended to comply with Rule 10b5-1 under the Exchange Act. Sales under these trading plans would not be permitted until the expiration of the lock-up agreements relating to the offering described above.
194
Table of Contents
MATERIAL U.S. FEDERAL INCOME TAX CONSEQUENCES TO NON-U.S. HOLDERS OF OUR COMMON STOCK
The following is a summary of the material U.S. federal income tax consequences to non-U.S. holders (as defined below) of the acquisition, ownership, and disposition of our common stock issued pursuant to this offering. This discussion is not a complete analysis of all potential U.S. federal income tax consequences relating thereto, does not address the potential application of the Medicare contribution tax on net investment income, the alternative minimum tax provisions of the Code, or the special tax accounting rules under Section 451(b) of the Code, and does not address any estate or gift tax consequences or any tax consequences arising under any state, local, or non-U.S. tax laws, or any U.S. federal tax laws other than U.S. federal income tax laws. This discussion is based on the Code and applicable Treasury Regulations promulgated thereunder, published rulings and administrative pronouncements of the Internal Revenue Service (IRS), and judicial decisions, all as in effect as of the date hereof. These authorities are subject to differing interpretations and may change, possibly retroactively, resulting in U.S. federal income tax consequences different from those discussed below. We have not requested a ruling from the IRS with respect to the statements made and the conclusions reached in the following summary, and there can be no assurance that the IRS or a court will agree with such statements and conclusions.
This discussion is limited to non-U.S. holders who purchase our common stock pursuant to this offering and who hold our common stock as a “capital asset” within the meaning of Section 1221 of the Code (generally, property held for investment). This discussion does not address all of the U.S. federal income tax consequences that may be relevant to a particular holder in light of such holder’s particular circumstances. This discussion also does not consider any specific facts or circumstances that may be relevant to holders subject to special rules under U.S. federal income tax laws, including but not limited to:
| ∎ | certain former citizens or long-term residents of the United States; |
| ∎ | “controlled foreign corporations” or “foreign controlled foreign corporations”; |
| ∎ | “passive foreign investment companies”; |
| ∎ | corporations that accumulate earnings to avoid U.S. federal income tax; |
| ∎ | banks, financial institutions, investment funds, insurance companies, brokers, dealers, or traders in securities or foreign currencies; |
| ∎ | tax-exempt organizations and governmental organizations; |
| ∎ | tax-qualified retirement plans; |
| ∎ | “qualified foreign pension funds” as defined in Section 897(l)(2) of the Code and entities all of the interests of which are held by qualified foreign pension funds; |
| ∎ | persons that own, or have owned, actually or constructively, more than 5% of our common stock at any time; |
| ∎ | persons who have elected to mark securities to market; |
| ∎ | persons holding our common stock as part of a hedging or conversion transaction or straddle, a constructive sale, or other risk reduction strategy or integrated investment; |
| ∎ | persons who acquire our common stock through the exercise of an employee option or otherwise as compensation; and |
| ∎ | corporations organized outside of the United States, any state thereof and the District of Columbia that are nonetheless treated as U.S. taxpayers for U.S. federal income tax purposes. |
If an entity or arrangement that is classified as a partnership for U.S. federal income tax purposes holds our common stock, the U.S. federal income tax treatment of the partnership and the partners thereof generally depends on the status of the partner and the activities of the partnership. Partnerships holding our common stock and the partners in such partnerships are urged to consult their tax advisors about the particular U.S. federal income tax consequences to them of holding and disposing of our common stock.
THIS DISCUSSION IS FOR INFORMATIONAL PURPOSES ONLY AND IS NOT TAX ADVICE. PROSPECTIVE INVESTORS SHOULD CONSULT THEIR TAX ADVISORS REGARDING THE PARTICULAR U.S. FEDERAL INCOME
195
Table of Contents
TAX CONSEQUENCES TO THEM OF ACQUIRING, OWNING, AND DISPOSING OF OUR COMMON STOCK, AS WELL AS ANY TAX CONSEQUENCES ARISING UNDER ANY STATE, LOCAL, OR NON-U.S. TAX LAWS AND ANY OTHER U.S. FEDERAL TAX LAWS.
Definition of Non-U.S. Holder
For purposes of this discussion, the term “non-U.S. holder” means any beneficial owner of our common stock that is neither a “U.S. person” nor a partnership (including any entity or arrangement treated as a partnership) for U.S. federal income tax purposes. A U.S. person is any person that, for U.S. federal income tax purposes, is or is treated as any of the following:
| ∎ | an individual who is a citizen or resident of the United States; |
| ∎ | a corporation (or entity treated as a corporation for U.S. federal income tax purposes) created or organized under the laws of the United States, any state thereof, or the District of Columbia; |
| ∎ | an estate, the income of which is subject to U.S. federal income tax regardless of its source; or |
| ∎ | a trust (i) whose administration is subject to the primary supervision of a U.S. court and which has one or more U.S. persons who have the authority to control all substantial decisions of the trust or (ii) that has a valid election in effect under applicable Treasury Regulations to be treated as a U.S. person. |
Distributions on our Common Stock
As described in the section titled “Dividend Policy,” we have not paid cash dividends on our common stock and do not anticipate paying cash dividends on our common stock for the foreseeable future. However, if we make cash or other property distributions on our common stock, such distributions will constitute dividends for U.S. federal income tax purposes to the extent paid from our current or accumulated earnings and profits, as determined under U.S. federal income tax principles. Amounts that exceed our current and accumulated earnings and profits and, therefore, are not treated as dividends for U.S. federal income tax purposes will constitute a return of capital and will first be applied against and reduce a holder’s tax basis in our common stock, but not below zero. Any amount distributed in excess of basis will be treated as gain realized on the sale or other disposition of our common stock and will be treated as described under “—Gain on Disposition of our Common Stock” below.
Subject to the discussions below regarding effectively connected income, backup withholding, and Sections 1471 through 1474 of the Code (commonly referred to as the Foreign Account Tax Compliance Act (FATCA)), dividends paid to a non-U.S. holder generally will be subject to U.S. federal withholding tax at a rate of 30% of the gross amount of the dividends or such lower rate specified by an applicable income tax treaty. To receive the benefit of a reduced treaty rate, a non-U.S. holder must furnish us or our paying agent with a valid IRS Form W-8BEN or IRS Form W-8BEN-E (or applicable successor form) and satisfy applicable certification and other requirements. This certification must be provided to us or our paying agent before the payment of dividends and must be updated periodically. If the non-U.S. holder holds the stock through a financial institution or other agent acting on the non-U.S. holder’s behalf, the non-U.S. holder will be required to provide appropriate documentation to the agent, which then will be required to provide certification to us or our paying agent, either directly or through other intermediaries.
If a non-U.S. holder holds our common stock in connection with the conduct of a trade or business in the United States, and dividends paid on our common stock are effectively connected with such holder’s U.S. trade or business (and, if required by an applicable tax treaty, are attributable to such holder’s permanent establishment in the United States), the non-U.S. holder generally will be exempt from U.S. federal withholding tax, provided such non-U.S. holder complies with applicable certification and disclosure requirements. To claim the exemption, the non-U.S. holder generally must furnish a valid IRS Form W-8ECI (or applicable successor form) to the applicable withholding agent. However, any such effectively connected dividends paid on our common stock generally will be subject to U.S. federal income tax on a net income basis at the regular U.S. federal income tax rates in the same manner as if such holder were a resident of the United States. A non-U.S. holder that is a foreign corporation also may be subject to an additional branch profits tax equal to 30% (or such lower rate specified by an applicable income tax treaty) of its effectively connected earnings and profits for the taxable year, as adjusted for certain items. Non-U.S. holders should consult their tax advisors regarding any applicable income tax treaties that may provide for different rules.
196
Table of Contents
Non-U.S. holders that do not provide the required certification on a timely basis, but that qualify for a reduced treaty rate, may obtain a refund of any excess amounts withheld by timely filing an appropriate claim for refund with the IRS.
Gain on Disposition of our Common Stock
Subject to the discussions below regarding backup withholding and FATCA, a non-U.S. holder generally will not be subject to U.S. federal income tax on any gain realized on the sale or other taxable disposition of our common stock, unless:
| ∎ | the gain is effectively connected with the non-U.S. holder’s conduct of a trade or business in the United States, and if required by an applicable income tax treaty, is attributable to a permanent establishment maintained by the non-U.S. holder in the United States; |
| ∎ | the non-U.S. holder is a nonresident alien individual present in the United States for 183 days or more during the taxable year of the disposition, and certain other requirements are met; or |
| ∎ | our common stock constitutes a United States real property interest (USRPI), by reason of our status as a United States real property holding corporation (USRPHC), for U.S. federal income tax purposes at any time within the shorter of the five-year period preceding the disposition or the non-U.S. holder’s holding period for our common stock, and our common stock is not regularly traded on an established securities market as defined by applicable Treasury Regulations. |
The determination of whether we are a USRPHC depends on the fair market value of our USRPIs relative to the fair market value of worldwide real property interests and our other assets used or held for use in a trade or business. We believe that we are not currently, and we do not anticipate becoming, a USRPHC for U.S. federal income tax purposes, although there can be no assurance we will not become a USRPHC in the future. Even if we are or were to become a USRPHC, gain arising from the sale or other taxable disposition of our common stock by a non-U.S. holder will not be subject to U.S. federal income tax if our common stock is “regularly traded” (as defined by applicable Treasury Regulations) on an established securities market. Prospective investors are encouraged to consult their own tax advisors regarding the possible consequences to them if we are, or were to become, a USRPHC.
Gain described in the first bullet point above generally will be subject to U.S. federal income tax on a net income basis at the regular U.S. federal income tax rates in the same manner as if such holder were a resident of the United States. A non-U.S. holder that is a foreign corporation also may be subject to an additional branch profits tax equal to 30% (unless an applicable income tax treaty provides for different treatment) of its effectively connected earnings and profits for the taxable year, as adjusted for certain items. A non-U.S. holder described in the second bullet point above will be subject to U.S. federal income tax at a flat 30% rate (or such lower rate specified by an applicable income tax treaty) on gain realized upon the sale or other taxable disposition of our common stock, but such gain may be offset by certain U.S.-source capital losses (even though the individual is not considered a resident of the United States), provided that the non-U.S. holder has timely filed U.S. federal income tax returns with respect to such losses. Non-U.S. holders should consult their tax advisors regarding any applicable income tax treaties that may provide for different rules.
Information Reporting and Backup Withholding
Annual reports are required to be filed with the IRS and provided to each non-U.S. holder indicating the amount of distributions on our common stock paid to such holder and the amount of any tax withheld with respect to those distributions. These information reporting requirements apply regardless of whether such distributions constitute dividends and even if no withholding was required. This information also may be made available under a specific treaty or agreement with the tax authorities in the country in which the non-U.S. holder resides or is established. Backup withholding, currently at a 24% rate, generally will not apply to payments to a non-U.S. holder of dividends on, or the gross proceeds of a disposition of, our common stock provided the non-U.S. holder furnishes the required certification for its non-U.S. status, such as by providing a valid IRS Form W-8BEN, IRS Form W-8BEN-E, or IRS Form W-8ECI, or certain other requirements are met. Backup withholding may apply if the applicable withholding agent has actual knowledge, or reason to know, that the holder is a U.S. person who is not an exempt recipient.
197
Table of Contents
Backup withholding is not an additional tax. If any amount is withheld under the backup withholding rules, the non-U.S. holder should consult with a U.S. tax advisor regarding the possibility of and procedure for obtaining a refund or a credit against the non-U.S. holder’s U.S. federal income tax liability, if any.
FATCA
FATCA imposes a U.S. federal withholding tax of 30% on certain payments made to a “foreign financial institution” (as specially defined under these rules) unless such institution enters into an agreement with the U.S. government to withhold on certain payments and to collect and provide to the U.S. tax authorities substantial information regarding certain U.S. account holders of such institution (which includes certain equity and debt holders of such institution, as well as certain account holders that are foreign entities with U.S. owners) or an exemption applies. FATCA also generally will impose a U.S. federal withholding tax of 30% on certain payments made to a non-financial foreign entity unless such entity provides the withholding agent a certification identifying certain direct and indirect U.S. owners of the entity or an exemption applies. An intergovernmental agreement between the United States and an applicable foreign country may modify these requirements. Under certain circumstances, a non-U.S. holder might be eligible for refunds or credits of such taxes. FATCA currently applies to certain U.S. source payments such as dividends paid on our common stock. Under applicable Treasury Regulations and administrative guidance, withholding under FATCA would have applied to payments of gross proceeds from the sale or other disposition of stock, but under proposed Treasury Regulations (the preamble to which specifies that taxpayers are permitted to rely on such proposed regulations pending finalization), no withholding would apply with respect to payments of gross proceeds.
Prospective investors are encouraged to consult with their own tax advisors regarding the possible implications of the FATCA rules on their investment in our common stock.
198
Table of Contents
Subject to the terms and conditions set forth in the underwriting agreement, dated , 2026, among us and Jefferies LLC, TD Securities (USA) LLC, Evercore Group L.L.C., and Cantor Fitzgerald & Co., as the representatives of the underwriters named below and the joint book-running managers of this offering, we have agreed to sell to the underwriters, and each of the underwriters has agreed, severally and not jointly, to purchase from us, the respective number of shares of common stock shown opposite its name below:
| UNDERWRITER |
NUMBER OF SHARES |
|||
| Jefferies LLC |
||||
| TD Securities (USA) LLC |
||||
| Evercore Group L.L.C. |
||||
| Cantor Fitzgerald & Co. |
||||
|
|
|
|||
| Total |
21,666,667 | |||
|
|
|
|||
The underwriting agreement provides that the obligations of the several underwriters are subject to certain conditions precedent such as the receipt by the underwriters of officers’ certificates and legal opinions and approval of certain legal matters by their counsel. The underwriting agreement provides that the underwriters will purchase all of the shares of common stock if any of them are purchased. If an underwriter defaults, the underwriting agreement provides that the purchase commitments of the nondefaulting underwriters may be increased or the underwriting agreement may be terminated. We have agreed to indemnify the underwriters and certain of their controlling persons against certain liabilities, including liabilities under the Securities Act, and to contribute to payments that the underwriters may be required to make in respect of those liabilities.
The underwriters have advised us that, following the completion of this offering, they currently intend to make a market in the common stock as permitted by applicable laws and regulations. However, the underwriters are not obligated to do so, and the underwriters may discontinue any market-making activities at any time without notice in their sole discretion. Accordingly, no assurance can be given as to the liquidity of the trading market for the common stock, that you will be able to sell any of the common stock held by you at a particular time or that the prices that you receive when you sell will be favorable.
The underwriters are offering the shares of common stock subject to their acceptance of the shares of common stock from us and subject to prior sale. The underwriters reserve the right to withdraw, cancel or modify offers to the public and to reject orders in whole or in part.
Commission and Expenses
The underwriters have advised us that they propose to offer the shares of common stock to the public at the initial public offering price set forth on the cover page of this prospectus and to certain dealers, which may include the underwriters, at that price less a concession not in excess of $ per share of common stock. After the offering, the initial public offering price and concession to dealers may be reduced by the representatives. No such reduction will change the amount of proceeds to be received by us as set forth on the cover page of this prospectus.
199
Table of Contents
The following table shows the public offering price, the underwriting discounts, and commissions that we are to pay the underwriters and the proceeds, before expenses, to us in connection with this offering. Such amounts are shown assuming both no exercise and full exercise of the underwriters’ option to purchase additional shares.
| PER SHARE | TOTAL | |||||||||||||||
| WITHOUT OPTION TO PURCHASE ADDITIONAL SHARES |
WITH OPTION TO PURCHASE ADDITIONAL SHARES |
WITHOUT OPTION TO PURCHASE ADDITIONAL SHARES |
WITH OPTION TO PURCHASE ADDITIONAL SHARES |
|||||||||||||
| Public offering price |
$ | $ | $ | $ | ||||||||||||
| Underwriting discounts and commissions paid by us |
$ | $ | $ | $ | ||||||||||||
| Proceeds to us, before expenses |
$ | $ | $ | $ | ||||||||||||
We estimate expenses payable by us in connection with this offering, other than the underwriting discounts and commissions referred to above, will be approximately $5,650,000. We have agreed to reimburse the underwriters for certain of their expenses, up to $40,000.
Determination of Offering Price
Prior to this offering, there has not been a public market for our common stock. Consequently, the initial public offering price for our common stock will be determined by negotiations between us and the representatives. Among the factors to be considered in these negotiations will be prevailing market conditions, our financial information, market valuations of other companies that we and the underwriters believe to be comparable to us, estimates of our business potential, the present state of our development, and other factors deemed relevant.
We offer no assurances that the initial public offering price will correspond to the price at which the common stock will trade in the public market subsequent to the offering or that an active trading market for the common stock will develop and continue after the offering.
Listing
We have applied to have our common stock listed on the Nasdaq Global Select Market under the trading symbol “ETRA.”
Stamp Taxes
If you purchase shares of common stock offered in this prospectus, you may be required to pay stamp taxes and other charges under the laws and practices of the country of purchase, in addition to the offering price listed on the cover page of this prospectus.
Option to Purchase Additional Shares
We have granted to the underwriters an option, exercisable for 30 days from the date of this prospectus, to purchase, from time to time, in whole or in part, up to an aggregate of 3,250,000 shares from us at the public offering price set forth on the cover page of this prospectus, less underwriting discounts and commissions. If the underwriters exercise this option, each underwriter will be obligated, subject to specified conditions, to purchase a number of additional shares proportionate to that underwriter’s initial purchase commitment as indicated in the table above. This option may be exercised only if the underwriters sell more shares than the total number set forth on the cover page of this prospectus.
No Sales of Similar Securities
We, our officers, directors, and holders of all or substantially all of our outstanding capital stock and other securities have agreed, subject to specified exceptions, not to directly or indirectly:
| ∎ | sell, offer to sell, contract or grant any option to sell (including any short sale), pledge, transfer, establish an open “put equivalent position” within the meaning of Rule 16a-1(h) under the Exchange Act, or otherwise |
200
Table of Contents
| dispose of any shares of common stock, options, or warrants to acquire shares of common stock, or securities exchangeable or exercisable for or convertible into shares of common stock currently or hereafter owned either of record or beneficially, or |
| ∎ | publicly announce an intention to do any of the foregoing for a period of 180 days after the date of this prospectus without the prior written consent of Jefferies LLC and TD Securities (USA) LLC in the case of our officers, directors and holders of our outstanding capital stock, and, in our case, without the prior written consent of Jefferies LLC, TD Securities (USA) LLC and Evercore Group L.L.C. |
This restriction terminates after the close of trading of the common stock on and including the 180th day after the date of this prospectus.
Notwithstanding the foregoing, the securityholder may transfer shares of common stock, or securities exchangeable or exercisable for or convertible into shares of common stock:
(i) by a bona fide gift or gifts or charitable contributions, or for bona fide estate planning purposes;
(ii) by will or intestate succession upon the death of the securityholder;
(iii) to any trust for the direct or indirect benefit of the securityholder or a family member of the securityholder;
(iv) to any family member of the securityholder;
(v) by operation of law pursuant to a court order or a settlement agreement related to the distribution of assets in connection with the dissolution of a marriage or civil union;
(vi) to any corporation, partnership, limited liability company, or other entity all of the beneficial ownership interests of which, in each case, are held by the securityholder;
(vii) in connection with a distribution or other transfer by a partnership to its partners or former partners or by a limited liability company to its members (including the contemplated distribution by Electra LLC of shares of our Series A redeemable convertible preferred stock to its equityholders prior to the closing of this offering) or retired members or by a corporation to its stockholders or former stockholders or to any wholly-owned subsidiary of such corporation;
(viii) to any affiliate of the securityholder, including investment funds, or other entities under common control or management that are affiliates of the securityholder;
(ix) if the securityholder is not one of our officers or directors, in connection with transactions relating to our shares of common stock acquired in this offering or in open market transactions on or after the completion of this offering;
(x) to us in connection with the exercise, vesting, exchange, or settlement of options, warrants, or other rights to acquire shares of common stock, including any security convertible into, exchangeable for, or that represent the right to receive shares of common stock, in accordance with their terms (including the vesting or settlement of restricted stock units and including, in each case, by way of net exercise and/or to cover withholding tax obligations in connection with such exercise, vesting, exchange, or settlement) pursuant to an employee benefit plan, option, warrant or other right disclosed in this prospectus;
(xi) to us in connection with (A) the termination of the securityholder’s employment with us or (B) any agreements under which we have the option to repurchase such shares;
(xii) upon exercise by the securityholder of any option to purchase any shares of common stock or the settlement of restricted stock units or other equity awards pursuant to any stock incentive plan or stock purchase plan disclosed in this prospectus, provided that the underlying shares of common stock shall continue to be subject to the restrictions on transfer set forth in the lock-up agreement;
(xiii) in connection with the conversion of any outstanding preferred stock into shares of common stock, provided that any such shares of common stock received upon such conversion shall be subject to the restrictions on transfer set forth in the lock-up agreement; and
(xiv) pursuant to a bona fide third-party tender offer for our securities, merger, consolidation, or other similar transaction made to all holders of our securities involving a change of control, which transaction is approved by our board of directors, provided that it shall be a condition of the transfer that if the tender offer, merger, consolidation, or other such transaction is not completed, the securityholder’s securities, which are subject to the lock-up agreement shall remain subject to the restrictions set forth in such lock-up agreement;
provided that, in the case of (a) clauses (i)-(viii), that (x) such transfer shall not involve a disposition for value and (y) any such transferee executes and delivers to the representatives a lock-up agreement; (b) clauses (vi) through (ix),
201
Table of Contents
that no public disclosure nor any filing by any party under Section 16(a) of the Exchange Act shall be required or shall be made voluntarily in connection with such transfer; and (c) clauses (i) through (v) and (x) through (xiii), that (x) no public disclosure nor any filing by any party under Section 16(a) of the Exchange Act shall be made voluntarily in connection with such transfer and (y) any required filing by any party under Section 16(a) of the Exchange Act or any other public filing, report, or announcement legally required during the lock-up period shall include a statement in such report indicating the circumstances in connection with such transfer.
In addition, the foregoing restrictions shall not apply to the establishment or amendment of any contract, instruction, or plan (Plan) that satisfies all of the requirements of Rule 10b5-1(c)(1)(i)(B) under the Exchange Act; provided that no sales of a securityholder’s shares of common stock shall be made pursuant to such Plan prior to the expiration of the lock-up period, and such Plan may only be established or amended if any required public disclosure, announcement or filing under the Exchange Act made by us or any person regarding the establishment or amendment of such Plan during the lock-up period shall include a statement that the securityholder is not permitted to transfer, sell, or otherwise dispose of securities under such Plan during the lock-up period in contravention of the lock-up agreement, and no announcement or filing is made voluntarily, by the securityholder, us, or any other person, regarding the establishment or amendment of such Plan, prior to the expiration of the lock-up period.
Jefferies LLC and TD Securities (USA) LLC may, in their sole discretion and at any time or from time to time before the termination of the 180-day period release all or any portion of the securities subject to lock-up agreements. There are no existing agreements between the underwriters and any of our stockholders who will execute a lock-up agreement, providing consent to the sale of shares prior to the expiration of the lock-up period.
Stabilization
The underwriters have advised us that, pursuant to Regulation M under the Exchange Act, certain persons participating in the offering may engage in short sale transactions, stabilizing transactions, syndicate covering transactions, or the imposition of penalty bids in connection with this offering. These activities may have the effect of stabilizing or maintaining the market price of the common stock at a level above that which might otherwise prevail in the open market. Establishing short sales positions may involve either “covered” short sales or “naked” short sales.
“Covered” short sales are sales made in an amount not greater than the underwriters’ option to purchase additional shares of our common stock in this offering. The underwriters may close out any covered short position by either exercising their option to purchase additional shares of our common stock or purchasing shares of our common stock in the open market. In determining the source of shares to close out the covered short position, the underwriters will consider, among other things, the price of shares available for purchase in the open market as compared to the price at which they may purchase shares through the option to purchase additional shares.
“Naked” short sales are sales in excess of the option to purchase additional shares of our common stock. The underwriters must close out any naked short position by purchasing shares in the open market. A naked short position is more likely to be created if the underwriters are concerned that there may be downward pressure on the price of the shares of our common stock in the open market after pricing that could adversely affect investors who purchase in this offering.
A stabilizing bid is a bid for the purchase of shares of common stock on behalf of the underwriters for the purpose of fixing or maintaining the price of the common stock. A syndicate covering transaction is the bid for or the purchase of shares of common stock on behalf of the underwriters to reduce a short position incurred by the underwriters in connection with the offering. Similar to other purchase transactions, the underwriter’s purchases to cover the syndicate short sales may have the effect of raising or maintaining the market price of our common stock or preventing or retarding a decline in the market price of our common stock. As a result, the price of our common stock may be higher than the price that might otherwise exist in the open market. A penalty bid is an arrangement permitting the underwriters to reclaim the selling concession otherwise accruing to a syndicate member in connection with the offering if the common stock originally sold by such syndicate member are purchased in a syndicate covering transaction and therefore have not been effectively placed by such syndicate member.
202
Table of Contents
Neither we nor any of the underwriters make any representation or prediction as to the direction or magnitude of any effect that the transactions described above may have on the price of our common stock. The underwriters are not obligated to engage in these activities and, if commenced, any of the activities may be discontinued at any time.
The underwriters may also engage in passive market making transactions in our common stock on the Nasdaq Global Select Market in accordance with Rule 103 of Regulation M during a period before the commencement of offers or sales of shares of our common stock in this offering and extending through the completion of distribution. A passive market maker must display its bid at a price not in excess of the highest independent bid of that security. However, if all independent bids are lowered below the passive market maker’s bid, that bid must then be lowered when specified purchase limits are exceeded.
Electronic Distribution
A prospectus in electronic format may be made available by e-mail or on the web sites or through online services maintained by one or more of the underwriters or their affiliates. In those cases, prospective investors may view offering terms online and may be allowed to place orders online. The underwriters may agree with us to allocate a specific number of shares of common stock for sale to online brokerage account holders. Any such allocation for online distributions will be made by the underwriters on the same basis as other allocations. Other than the prospectus in electronic format, the information on the underwriters’ web sites and any information contained in any other web site maintained by any of the underwriters is not part of this prospectus, has not been approved and/or endorsed by us or the underwriters and should not be relied upon by investors.
Other Activities and Relationships
The underwriters and certain of their respective affiliates are full service financial institutions engaged in various activities, which may include securities trading, commercial and investment banking, financial advisory, investment management, investment research, principal investment, hedging, financing, and brokerage activities. The underwriters and certain of their respective affiliates have, from time to time, performed, and may in the future perform, various commercial and investment banking and financial advisory services for us and our affiliates, for which they received or will receive customary fees and expenses.
In the ordinary course of their various business activities, the underwriters and certain of their respective affiliates may make or hold a broad array of investments and actively trade debt and equity securities (or related derivative securities) and financial instruments (including bank loans) for their own account and for the accounts of their customers, and such investment and securities activities may involve securities and/or instruments issued by us and our affiliates. If the underwriters or their respective affiliates have a lending relationship with us, they routinely hedge their credit exposure to us consistent with their customary risk management policies. The underwriters and their respective affiliates may hedge such exposure by entering into transactions which consist of either the purchase of credit default swaps or the creation of short positions in our securities or the securities of our affiliates, including potentially the common stock offered hereby. Any such short positions could adversely affect future trading prices of the common stock offered hereby. The underwriters and certain of their respective affiliates may also communicate independent investment recommendations, market color, or trading ideas and/or publish or express independent research views in respect of such securities or instruments and may at any time hold, or recommend to clients that they acquire, long and/or short positions in such securities and instruments.
Disclaimers About Non-U.S. Jurisdictions
Canada
(A) Resale Restrictions
The distribution of shares of common stock in Canada is being made only in the provinces of Ontario, Quebec, Alberta, and British Columbia on a private placement basis exempt from the requirement that we prepare and file a prospectus with the securities regulatory authorities in each province where trades of these securities are made. Any resale of the shares of common stock in Canada must be made under applicable securities laws which may vary depending on the relevant jurisdiction, and which may require resales to be made under available statutory exemptions or under a discretionary exemption granted by the applicable Canadian securities regulatory authority. Purchasers are advised to seek legal advice prior to any resale of the shares of common stock.
203
Table of Contents
(B) Representations of Canadian Purchasers
By purchasing shares of common stock in Canada and accepting delivery of a purchase confirmation, a purchaser is representing to us and the dealer from whom the purchase confirmation is received that:
| ∎ | the purchaser is entitled under applicable provincial securities laws to purchase the shares of common stock without the benefit of a prospectus qualified under those securities laws as it is an “accredited investor” as defined under National Instrument 45-106 - Prospectus Exemptions or Section 73.3(1) of the Securities Act (Ontario), as applicable, |
| ∎ | the purchaser is a “permitted client” as defined in National Instrument 31-103 - Registration Requirements, Exemptions and Ongoing Registrant Obligations, |
| ∎ | where required by law, the purchaser is purchasing as principal and not as agent, and |
| ∎ | the purchaser has reviewed the text above under Resale Restrictions. |
(C) Conflicts of Interest
Canadian purchasers are hereby notified that certain of the underwriters are relying on the exemption set out in section 3A.3 or 3A.4, if applicable, of National Instrument 33-105—Underwriting Conflicts from having to provide certain conflict of interest disclosure in this prospectus.
(D) Statutory Rights of Action
Securities legislation in certain provinces or territories of Canada may provide a purchaser with remedies for rescission or damages if the prospectus (including any amendment thereto) such as this prospectus contains a misrepresentation, provided that the remedies for rescission or damages are exercised by the purchaser within the time limit prescribed by the securities legislation of the purchaser’s province or territory. The purchaser of these securities in Canada should refer to any applicable provisions of the securities legislation of the purchaser’s province or territory for particulars of these rights or consult with a legal advisor.
(E) Enforcement of Legal Rights
All of our directors and officers as well as the experts named herein may be located outside of Canada and, as a result, it may not be possible for Canadian purchasers to effect service of process within Canada upon us or those persons. All or a substantial portion of our assets and the assets of those persons may be located outside of Canada and, as a result, it may not be possible to satisfy a judgment against us or those persons in Canada or to enforce a judgment obtained in Canadian courts against us or those persons outside of Canada.
(F) Taxation and Eligibility for Investment
Canadian purchasers of shares of common stock should consult their own legal and tax advisors with respect to the tax consequences of an investment in the shares of common stock in their particular circumstances and about the eligibility of the shares of common stock for investment by the purchaser under relevant Canadian legislation.
(G) Language of Documents
The purchaser confirms its express wish and that it has requested that this document, all documents evidencing or relating to the sale of the securities described herein and all other related documents be drawn up exclusively in the English language. L’acquéreur confirme sa volonté expresse et qu’il a demandé que le présent document, tous les documents attestant de la vente des titres décrits dans le présent document ou s’y rapportant ainsi que tous les autres documents s’y rattachant soient rédigés exclusivement en langue anglaise.
Australia
This prospectus is not a disclosure document for the purposes of Australia’s Corporations Act 2001 (Cth) of Australia (the Corporations Act), has not been lodged with the Australian Securities & Investments Commission and is only directed to the categories of exempt persons set out below. Accordingly, if you receive this prospectus in Australia:
You confirm and warrant that you are either:
| ∎ | a “sophisticated investor” under section 708(8)(a) or (b) of the Corporations Act; |
| ∎ | a “sophisticated investor” under section 708(8)(c) or (d) of the Corporations Act and that you have provided an accountant’s certificate to the Company which complies with the requirements of section 708(8)(c)(i) or (ii) of the Corporations Act and related regulations before the offer has been made; |
204
Table of Contents
| ∎ | a person associated with the Company under Section 708(12) of the Corporations Act; or |
| ∎ | a “professional investor” within the meaning of section 708(11)(a) or (b) of the Corporations Act. |
To the extent that you are unable to confirm or warrant that you are an exempt sophisticated investor, associated person or professional investor under the Corporations Act any offer made to you under this prospectus is void and incapable of acceptance.
You warrant and agree that you will not offer any of the securities issued to you pursuant to this prospectus for resale in Australia within 12 months of those securities being issued unless any such resale offer is exempt from the requirement to issue a disclosure document under section 708 of the Corporations Act.
European Economic Area
In relation to each Member State of the European Economic Area (each, a Relevant State), no shares have been offered or will be offered pursuant to the offering to the public in that Relevant State prior to the publication of a prospectus in relation to the shares of common stock which has been approved by the competent authority in that Relevant State or, where appropriate, approved in another Relevant State and notified to the competent authority in that Relevant State, all in accordance with the Prospectus Regulation, except that the shares of common stock may be offered to the public in that Relevant State at any time.
| (a) | to any legal entity which is a “qualified investor” as defined under Article 2 of the Prospectus Regulation; |
| (b) | to fewer than 150 natural or legal persons (other than qualified investors as defined under Article 2 of the Prospectus Regulation), subject to obtaining the prior consent of representatives for any such offer; or |
| (c) | in any other circumstances falling within Article 1(4) of the Prospectus Regulation, |
provided that no such offer of the shares of common stock shall require us or any of the representatives to publish a prospectus pursuant to Article 3 of the Prospectus Regulation, supplement a prospectus pursuant to Article 23 of the Prospectus Regulation or publish an Annex IX document pursuant to Article 1(4) of the Prospectus Regulation.
For the purposes of this provision, the expression “offer to the public” in relation to the shares of common stock in any Relevant State means the communication in any form and by any means of sufficient information on the terms of the offer and any shares to be offered so as to enable an investor to decide to purchase or subscribe for any shares, and the expression “Prospectus Regulation” means Regulation (EU) 2017/1129.
Hong Kong
No shares of common stock have been offered or sold, and no shares of common stock may be offered or sold, in Hong Kong, by means of any document, other than to persons whose ordinary business is to buy or sell shares or debentures, whether as principal or agent; or to “professional investors” as defined in the Securities and Futures Ordinance (Cap. 571) of Hong Kong (the SFO), and any rules made under that Ordinance; or in other circumstances which do not result in the document being a “prospectus” as defined in the Companies Ordinance (Cap. 32) of Hong Kong (the CO), or which do not constitute an offer or invitation to the public for the purpose of the CO or the SFO. No document, invitation, or advertisement relating to the shares of common stock has been issued or may be issued or may be in the possession of any person for the purpose of issue (in each case whether in Hong Kong or elsewhere), which is directed at, or the contents of which are likely to be accessed or read by, the public of Hong Kong (except if permitted under the securities laws of Hong Kong) other than with respect to shares of common stock which are or are intended to be disposed of only to persons outside Hong Kong or only to “professional investors” as defined in the SFO and any rules made under that Ordinance.
This prospectus has not been registered with the Registrar of Companies in Hong Kong. Accordingly, this prospectus may not be issued, circulated, or distributed in Hong Kong, and the shares of common stock may not be offered for subscription to members of the public in Hong Kong. Each person acquiring the shares of common stock will be required, and is deemed by the acquisition of the shares of common stock, to confirm that he is aware of the restriction on offers of the shares of common stock described in this prospectus and the relevant offering documents and that he is not acquiring, and has not been offered any shares of common stock in circumstances that contravene any such restrictions.
205
Table of Contents
Israel
This prospectus does not constitute a prospectus under the Israeli Securities Law, 5728-1968 (the Securities Law), and has not been filed with or approved by the Israel Securities Authority. In Israel, this prospectus is being distributed only to, and is directed only at, and any offer of the shares of common stock is directed only at, (i) a limited number of persons in accordance with the Israeli Securities Law and (ii) investors listed in the first addendum (the Addendum), to the Israeli Securities Law, consisting primarily of joint investment in trust funds, provident funds, insurance companies, banks, portfolio managers, investment advisors, members of the Tel Aviv Stock Exchange, underwriters, venture capital funds, entities with equity in excess of NIS 50 million, and “qualified individuals,” each as defined in the Addendum (as it may be amended from time to time), collectively referred to as qualified investors (in each case, purchasing for their own account or, where permitted under the Addendum, for the accounts of their clients who are investors listed in the Addendum). Qualified investors are required to submit written confirmation that they fall within the scope of the Addendum, are aware of the meaning of same, and agree to it.
Japan
The offering has not been and will not be registered under the Financial Instruments and Exchange Law of Japan (Law No. 25 of 1948 of Japan, as amended) (the FIEL), and the underwriters will not offer or sell any securities, directly or indirectly, in Japan or to, or for the benefit of, any resident of Japan (which term as used herein means any person resident in Japan, including any corporation or other entity organized under the laws of Japan), or to others for re-offering or resale, directly or indirectly, in Japan or to, or for the benefit of, any resident of Japan, except pursuant to an exemption from the registration requirements of, and otherwise in compliance with, the FIEL and any other applicable laws, regulations, and ministerial guidelines of Japan.
Singapore
This prospectus has not been and will not be lodged or registered as a prospectus with the Monetary Authority of Singapore. Accordingly, this prospectus and any other document or material in connection with the offer or sale, or invitation for subscription or purchase, of the common stock may not be circulated or distributed, nor may the common stock be offered or sold, or be made the subject of an invitation for subscription or purchase, whether directly or indirectly, to persons in Singapore other than (i) to an institutional investor under Section 274 of the Securities and Futures Act, Chapter 289 of Singapore (the SFA), (ii) to a relevant person pursuant to Section 275(1), or any person pursuant to Section 275(1A), and in accordance with the conditions specified in Section 275, of the SFA, or (iii) otherwise pursuant to, and in accordance with the conditions of, any other applicable provision of the SFA.
Where the shares of common stock are subscribed or purchased under Section 275 of the SFA by a relevant person which is:
| (a) | a corporation (which is not an accredited investor (as defined in Section 4A of the SFA)) the sole business of which is to hold investments and the entire share capital of which is owned by one or more individuals, each of whom is an accredited investor; or |
| (b) | a trust (where the trustee is not an accredited investor) whose sole purpose is to hold investments and each beneficiary of the trust is an individual who is an accredited investor, |
securities (as defined in Section 239(1) of the SFA) of that corporation or the beneficiaries’ rights and interest (howsoever described) in that trust shall not be transferred within six months after that corporation or that trust has acquired the shares of common stock pursuant to an offer made under Section 275 of the SFA except:
| (i) | to an institutional investor or to a relevant person defined in Section 275(2) of the SFA, or to any person arising from an offer referred to in Section 275(1A) or Section 276(4)(i)(B) of the SFA; |
| (ii) | where no consideration is or will be given for the transfer; |
| (iii) | where the transfer is by operation of law; |
| (iv) | as specified in Section 276(7) of the SFA; or |
| (v) | as specified in Regulation 32 of the Securities and Futures (Offers of Investments) (Shares and Debentures) Regulations 2005 of Singapore. |
206
Table of Contents
Switzerland
The shares of common stock may not be publicly offered in Switzerland and will not be listed on the SIX Swiss Exchange (the SIX), or on any other stock exchange or regulated trading facility in Switzerland. This prospectus has been prepared without regard to the disclosure standards for issuance prospectuses under art. 652a or art. 1156 of the Swiss Code of Obligations or the disclosure standards for listing prospectuses under art. 27 ff. of the SIX Listing Rules or the listing rules of any other stock exchange or regulated trading facility in Switzerland. Neither this prospectus nor any other offering or marketing material relating to the securities or the offering may be publicly distributed or otherwise made publicly available in Switzerland.
Neither this prospectus nor any other offering or marketing material relating to the offering, us or the securities have been or will be filed with or approved by any Swiss regulatory authority. In particular, this prospectus will not be filed with, and the offer of securities will not be supervised by, the Swiss Financial Market Supervisory Authority (the FINMA), and the offer of securities has not been and will not be authorized under the Swiss Federal Act on Collective Investment Schemes (the CISA). The investor protection afforded to acquirers of interests in collective investment schemes under the CISA does not extend to acquirers of securities.
United Kingdom
No shares of common stock have been offered or will be offered pursuant to the offering to the public in the United Kingdom except that the shares of common stock may be offered to the public in the United Kingdom at any time:
| (a) | where the offer is conditional on the admission of the shares of common stock to trading on the London Stock Exchange plc’s main market (in reliance on the exception in paragraph 6(a) of Schedule 1 of the POATR); |
| (b) | to any qualified investor as defined under paragraph 15 of Schedule 1 of the POATR; |
| (c) | to fewer than 150 persons (other than qualified investors as defined under paragraph 15 of Schedule 1 of the POATR), subject to obtaining the prior consent of the representatives for any such offer; or |
| (d) | in any other circumstances falling within Part 1 of Schedule 1 of the POATR. |
For the purposes of this provision, the expression an “offer to the public” in relation to the shares of common stock in the United Kingdom means the communication to any person which presents sufficient information on: (a) the shares of common stock to be offered; and (b) the terms on which they are to be offered, to enable an investor to decide to buy or subscribe for the shares of common stock and the expression “POATR” means the Public Offers and Admissions to Trading Regulations 2024.
207
Table of Contents
Table of Contents
Ernst & Young LLP, independent registered public accounting firm, has audited our financial statements at December 31, 2025 and 2024, and for the years then ended, as set forth in their report (which contains an explanatory paragraph describing conditions that raise substantial doubt about the Company’s ability to continue as a going concern as described in Note 1 to the financial statements). We have included our financial statements in the prospectus and elsewhere in the registration statement in reliance on Ernst & Young LLP’s report, given on their authority as experts in accounting and auditing.
209
Table of Contents
WHERE YOU CAN FIND MORE INFORMATION
We have filed with the SEC a registration statement on Form S-1, including exhibits and schedules, under the Securities Act, with respect to the shares of our common stock being offered by this prospectus. This prospectus, which constitutes part of the registration statement, does not contain all of the information in the registration statement and its exhibits. For further information with respect to us and the common stock offered by this prospectus, we refer you to the registration statement and its exhibits. Statements contained in this prospectus as to the contents of any contract or any other document referred to are not necessarily complete, and in each instance, we refer you to the copy of the contract or other document filed as an exhibit to the registration statement. Each of these statements is qualified in all respects by this reference.
You can read our SEC filings, including the registration statement, over the internet at the SEC’s website at www.sec.gov.
Upon the closing of this offering, we will be subject to the information reporting requirements of the Exchange Act, and we will file reports, proxy statements, and other information with the SEC. These reports, proxy statements, and other information will be available for inspection at the website of the SEC referred to above. We also maintain a website at www.electra-therapeutics.com, at which, following the closing of this offering, you may access these materials free of charge as soon as reasonably practicable after they are electronically filed with, or furnished to, the SEC. Information contained in or accessible through our website is not a part of this prospectus, and the inclusion of our website address in this prospectus is an inactive textual reference only.
210
Table of Contents
Audited Financial Statements as of December 31, 2025 and 2024 and for the Years Ended December 31, 2025 and 2024
| PAGE | ||||
| Report of Independent Registered Public Accounting Firm (PCAOB ID: 42) |
F-2 | |||
| F-3 | ||||
| F-4 | ||||
| Statements of Redeemable Convertible Preferred Stock and Stockholders’ Deficit |
F-5 | |||
| F-6 | ||||
| F-7 | ||||
Unaudited Interim Condensed Financial Statements as of June 30, 2026 and for the Six Months Ended June 30, 2026 and 2025
| PAGE | ||||
| F-28 | ||||
| F-29 | ||||
| F-30 | ||||
| Condensed Statements of Redeemable Convertible Preferred Stock and Stockholders’ Deficit |
F-31 | |||
| F-32 | ||||
| F-33 | ||||
F-1
Table of Contents
Report of Independent Registered Public Accounting Firm
To the Stockholders and the Board of Directors of Electra Therapeutics, Inc.
Opinion on the Financial Statements
We have audited the accompanying balance sheets of Electra Therapeutics, Inc. (the Company) as of December 31, 2025 and 2024, the related statements of operations and comprehensive loss, redeemable convertible preferred stock and stockholders’ deficit and cash flows for the years then ended and the related notes (collectively referred to as the “financial statements”). In our opinion, the financial statements present fairly, in all material respects, the financial position of the Company at December 31, 2025 and 2024, and the results of its operations and its cash flows for the years then ended in conformity with U.S. generally accepted accounting principles.
The Company’s Ability to Continue as a Going Concern
The accompanying financial statements have been prepared assuming that the Company will continue as a going concern. As discussed in Note 1 to the financial statements, the Company has suffered recurring losses from operations, and has stated that substantial doubt exists about the Company’s ability to continue as a going concern. Management’s evaluation of the events and conditions and management’s plans regarding these matters are also described in Note 1. The financial statements do not include any adjustments that might result from the outcome of this uncertainty.
Basis for Opinion
These financial statements are the responsibility of the Company’s management. Our responsibility is to express an opinion on the Company’s financial statements based on our audits. We are a public accounting firm registered with the Public Company Accounting Oversight Board (United States) (PCAOB) and are required to be independent with respect to the Company in accordance with the U.S. federal securities laws and the applicable rules and regulations of the Securities and Exchange Commission and the PCAOB.
We conducted our audits in accordance with the standards of the PCAOB. Those standards require that we plan and perform the audit to obtain reasonable assurance about whether the financial statements are free of material misstatement, whether due to error or fraud. The Company is not required to have, nor were we engaged to perform, an audit of its internal control over financial reporting. As part of our audits, we are required to obtain an understanding of internal control over financial reporting but not for the purpose of expressing an opinion on the effectiveness of the Company’s internal control over financial reporting. Accordingly, we express no such opinion.
Our audits included performing procedures to assess the risks of material misstatement of the financial statements, whether due to error or fraud, and performing procedures that respond to those risks. Such procedures included examining, on a test basis, evidence regarding the amounts and disclosures in the financial statements. Our audits also included evaluating the accounting principles used and significant estimates made by management, as well as evaluating the overall presentation of the financial statements. We believe that our audits provide a reasonable basis for our opinion.
/s/ Ernst & Young LLP
We have served as the Company’s auditor since 2022.
San Mateo, California
June 26, 2026,
except for the seventh paragraph of Note 1, as to which the date is
September 14, 2026
F-2
Table of Contents
ELECTRA THERAPEUTICS, INC.
(In thousands, except share and per share amounts)
| DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Assets |
||||||||
| Current assets: |
||||||||
| Cash and cash equivalents |
$ | 33,008 | $ | 14,787 | ||||
| Marketable securities |
53,510 | 998 | ||||||
| Other receivables |
220 | 55 | ||||||
| Prepaid expenses |
8,748 | 3,580 | ||||||
|
|
|
|
|
|||||
| Total current assets |
95,486 | 19,420 | ||||||
| Property, plant, and equipment, net |
674 | — | ||||||
| Right-of-use assets |
2,110 | — | ||||||
| Other assets, non-current |
1,098 | — | ||||||
|
|
|
|
|
|||||
| Total assets |
$ | 99,368 | $ | 19,420 | ||||
|
|
|
|
|
|||||
| Liabilities, redeemable convertible preferred stock, and stockholders’ deficit |
||||||||
| Current liabilities: |
||||||||
| Accounts payable |
$ | 4,076 | $ | 1,348 | ||||
| Accounts payable – related party |
— | 187 | ||||||
| Accrued liabilities |
5,045 | 2,538 | ||||||
| Accrued liabilities – related party |
— | 122 | ||||||
| Operating lease liability |
605 | — | ||||||
|
|
|
|
|
|||||
| Total current liabilities |
9,726 | 4,195 | ||||||
| Operating lease liability, net of current portion |
1,482 | — | ||||||
|
|
|
|
|
|||||
| Total liabilities |
11,208 | 4,195 | ||||||
|
|
|
|
|
|||||
| Commitments and contingencies (Note 9) |
||||||||
| Series A redeemable convertible preferred stock: $0.0001 par value – 22,601,626 shares authorized, 15,065,741 issued and outstanding as of December 31, 2025 and 2024, (liquidation preference of $34,750 as of December 31, 2025) |
34,650 | 34,650 | ||||||
| Series B redeemable convertible preferred stock: $0.0001 par value – 9,309,492 shares authorized, 6,205,494 issued and outstanding as of December 31, 2025 and 2024, (liquidation preference of $84,019 as of December 31, 2025) |
83,836 | 83,836 | ||||||
| Series C redeemable convertible preferred stock: $0.001 par value – 27,203,243 shares and 0 shares authorized as of December 31, 2025 and 2024, respectively. 13,214,580 and 0 shares issued and outstanding as of December 31, 2025 and 2024, respectively, (liquidation preference of $134,188 as of December 31, 2025) |
133,116 | — | ||||||
| Stockholders’ deficit: |
||||||||
| Common stock: $0.0001 par value – 71,000,000 and 37,229,638 |
— | — | ||||||
| Additional paid-in-capital |
9,515 | 7,741 | ||||||
| Accumulated deficit |
(172,978 | ) | (111,003 | ) | ||||
| Accumulated other comprehensive income |
21 | 1 | ||||||
|
|
|
|
|
|||||
| Total stockholders’ deficit |
(163,442 | ) | (103,261 | ) | ||||
|
|
|
|
|
|||||
| Total liabilities, redeemable convertible preferred stock and stockholders’ deficit |
$ | 99,368 | $ | 19,420 | ||||
|
|
|
|
|
|||||
See accompanying notes to the financial statements.
F-3
Table of Contents
ELECTRA THERAPEUTICS, INC.
Statements of Operations and Comprehensive Loss
(In thousands except share and per share amounts)
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Operating expenses: |
||||||||
| Research and development |
$ | 50,387 | $ | 19,267 | ||||
| General and administrative |
10,390 | 5,241 | ||||||
| General and administrative – related party |
1,112 | 1,809 | ||||||
|
|
|
|
|
|||||
| Total operating expenses |
61,889 | 26,317 | ||||||
|
|
|
|
|
|||||
| Loss from operations |
(61,889 | ) | (26,317 | ) | ||||
|
|
|
|
|
|||||
| Other income (expense), net: |
||||||||
| Interest income |
1,362 | 1,524 | ||||||
| Interest expense – related party |
(1,433 | ) | — | |||||
| Other income (expense) |
(15 | ) | 7 | |||||
|
|
|
|
|
|||||
| Total other income (expense), net |
(86 | ) | 1,531 | |||||
|
|
|
|
|
|||||
| Net loss |
$ | (61,975 | ) | $ | (24,786 | ) | ||
|
|
|
|
|
|||||
| Other comprehensive income: |
||||||||
| Unrealized gain on marketable securities |
20 | — | ||||||
|
|
|
|
|
|||||
| Total comprehensive loss |
$ | (61,955 | ) | $ | (24,786 | ) | ||
|
|
|
|
|
|||||
| Net loss per share, basic and diluted |
$ | (2,113.10 | ) | $ | (845.10 | ) | ||
|
|
|
|
|
|||||
| Weighted-average common shares outstanding, basic and diluted |
29,329 | 29,329 | ||||||
|
|
|
|
|
|||||
See accompanying notes to the financial statements.
F-4
Table of Contents
ELECTRA THERAPEUTICS, INC.
Statements of Redeemable Convertible Preferred Stock and Stockholders’ Deficit
(In thousands, except share amounts)
| SERIES A REDEEMABLE CONVERTIBLE PREFERRED STOCK |
SERIES B REDEEMABLE CONVERTIBLE PREFERRED STOCK |
SERIES C REDEEMABLE CONVERTIBLE PREFERRED STOCK |
COMMON STOCK | ADDITIONAL PAID-IN CAPITAL |
ACCUMULATED DEFICIT |
OTHER COMPREHENSIVE INCOME |
TOTAL STOCKHOLDERS’ DEFICIT |
|||||||||||||||||||||||||||||||||||||||||
| SHARES | AMOUNT | SHARES | AMOUNT | SHARES | AMOUNT | SHARES | AMOUNT | |||||||||||||||||||||||||||||||||||||||||
| Balance as of December 31, 2023 |
15,065,741 | $ | 34,650 | 6,205,494 | $ | 83,836 | — | $ | — | 29,329 | $ | — | $ | 5,432 | $ | (86,217 | ) | $ | 1 | $ | (80,784 | ) | ||||||||||||||||||||||||||
| Stock-based compensation - Profits interest units |
— | — | — | — | — | — | — | — | 1,053 | — | — | 1,053 | ||||||||||||||||||||||||||||||||||||
| Stock-based compensation expense - options |
— | — | — | — | — | — | — | — | 1,256 | — | — | 1,256 | ||||||||||||||||||||||||||||||||||||
| Net loss |
— | — | — | — | — | — | — | — | — | (24,786 | ) | — | (24,786 | ) | ||||||||||||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
| Balance as of December 31, 2024 |
15,065,741 | $ | 34,650 | 6,205,494 | $ | 83,836 | — | $ | — | 29,329 | $ | — | $ | 7,741 | $ | (111,003 | ) | $ | 1 | $ | (103,261 | ) | ||||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
| Conversion of Notes into Series C redeemable convertible preferred stock – related party |
— | — | — | — | 4,080,265 | 41,433 | — | — | — | — | — | — | ||||||||||||||||||||||||||||||||||||
| Issuance of Series C redeemable convertible preferred stock, net of issuance costs |
— | — | — | — | 9,134,315 | 91,683 | — | — | — | — | — | — | ||||||||||||||||||||||||||||||||||||
| Stock-based compensation - Profits interest units |
— | — | — | — | — | — | — | — | 480 | — | — | 480 | ||||||||||||||||||||||||||||||||||||
| Stock-based compensation expense - options |
— | — | — | — | — | — | — | — | 1,294 | — | — | 1,294 | ||||||||||||||||||||||||||||||||||||
| Other comprehensive income |
— | — | — | — | — | — | — | — | — | — | 20 | 20 | ||||||||||||||||||||||||||||||||||||
| Net loss |
— | — | — | — | — | — | — | — | — | (61,975 | ) | — | (61,975 | ) | ||||||||||||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
| Balance as of December 31, 2025 |
15,065,741 | $ | 34,650 | 6,205,494 | $ | 83,836 | 13,214,580 | $ | 133,116 | 29,329 | $ | — | $ | 9,515 | $ | (172,978 | ) | $ | 21 | $ | (163,442 | ) | ||||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
See accompanying notes to the financial statements.
F-5
Table of Contents
ELECTRA THERAPEUTICS, INC.
(In thousands)
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Cash flows from operating activities |
||||||||
| Net loss |
$ | (61,975 | ) | $ | (24,786 | ) | ||
| Adjustments to reconcile net loss to net cash used in operating activities: |
||||||||
| Stock-based compensation expense – profits interest units and options |
1,774 | 2,309 | ||||||
| Depreciation and amortization |
61 | — | ||||||
| Non-cash interest expense – related party |
1,433 | — | ||||||
| Non-cash lease expense |
215 | — | ||||||
| Amortization of premiums and discounts of marketable securities, net |
(248 | ) | (722 | ) | ||||
| Changes in assets and liabilities: |
||||||||
| Other receivables |
(165 | ) | 38 | |||||
| Prepaid expenses |
(5,168 | ) | (3,259 | ) | ||||
| Operating lease liabilities |
(239 | ) | — | |||||
| Other assets |
(1,098 | ) | — | |||||
| Accounts payable |
2,728 | 1,081 | ||||||
| Accounts payable – related party |
(187 | ) | (983 | ) | ||||
| Accrued liabilities |
2,507 | 484 | ||||||
| Accrued liabilities – related party |
(122 | ) | (165 | ) | ||||
|
|
|
|
|
|||||
| Net cash used in operating activities |
(60,484 | ) | (26,003 | ) | ||||
|
|
|
|
|
|||||
| Cash flows from investing activities |
||||||||
| Proceeds from maturity of marketable securities |
1,000 | 34,500 | ||||||
| Purchase of equipment |
(733 | ) | — | |||||
| Purchases of marketable securities |
(53,245 | ) | (4,799 | ) | ||||
|
|
|
|
|
|||||
| Net cash provided by (used in) investing activities |
(52,978 | ) | 29,701 | |||||
|
|
|
|
|
|||||
| Cash flows from financing activities |
||||||||
| Proceeds from the issuance of convertible promissory notes – related party |
40,000 | — | ||||||
| Proceeds from the issuance of Series C redeemable convertible preferred stock, net of $1.1 million issuance costs |
91,683 | — | ||||||
|
|
|
|
|
|||||
| Net cash provided by financing activities |
131,683 | — | ||||||
|
|
|
|
|
|||||
| Net increase in cash and cash equivalents |
18,221 | 3,698 | ||||||
| Cash and cash equivalents at the beginning of the year |
14,787 | 11,089 | ||||||
|
|
|
|
|
|||||
| Cash and cash equivalents at the end of the year |
$ | 33,008 | $ | 14,787 | ||||
|
|
|
|
|
|||||
| Supplemental disclosures: |
||||||||
| Cash paid for taxes |
$ | 8 | $ | 27 | ||||
|
|
|
|
|
|||||
| Initial recognition of right-of-use asset in exchange for lease liability |
$ | 2,325 | $ | — | ||||
|
|
|
|
|
|||||
| Conversion of Notes into Series C redeemable convertible preferred stock |
$ | 40,000 | $ | — | ||||
|
|
|
|
|
|||||
| Conversion of non-cash interest expense – related party into Series C redeemable convertible preferred stock |
$ | 1,433 | $ | — | ||||
|
|
|
|
|
|||||
See accompanying notes to the financial statements.
F-6
Table of Contents
1. Organization and Description of Business
Description of Business
Electra Therapeutics, Inc. (“Electra”, or the “Company”) was incorporated in 2018 and is a late clinical-stage biopharmaceutical company focused on pioneering a new class of precision medicines for the treatment of immune-mediated diseases and cancer. The Company’s novel approach targets signal regulatory proteins (“SIRP”), a family of cell surface receptors whose expression is restricted to specific immune cell populations and increases upon activation, to enable selective depletion of disease-driving cells while preserving normal immune function. The Company’s lead product candidate is ipsoprubart, a novel pan-SIRP monoclonal antibody designed to selectively deplete pathological myeloid cells and T cells via binding to SIRPa/ß1/g. The Company is also advancing ELA822, a novel SIRPg-specific monoclonal antibody designed to selectively deplete activated T cells, with potential applications across chronic T cell-mediated immune and inflammatory diseases.
Liquidity and Going Concern
The Company has evaluated whether there are conditions and events which raise substantial doubt about its ability to continue as a going concern within one year after the date the financial statements are available to be issued. The accompanying financial statements are prepared in accordance with generally accepted accounting principles applicable to a going concern, which contemplates the realization of assets and the satisfaction of liabilities in the normal course of business.
As of December 31, 2025, the Company had assets of $99.4 million consisting primarily of cash, cash equivalents and marketable securities of $86.5 million with total liabilities of $11.2 million. The Company has incurred significant net operating losses from operations and had an accumulated deficit of approximately $173.0 million as of December 31, 2025. The Company incurred a net loss of approximately $62.0 million and used approximately $60.5 million of cash in operating activities for the year ended December 31, 2025. The Company had no debt obligation outstanding as of December 31, 2025. Additionally, the Company anticipates that its net operating losses and negative operating cash flows will continue to increase for the foreseeable future as it continues to expand its research and development programs. The Company expects that its cash, cash equivalents, and marketable securities as of December 31, 2025 together with the proceeds from the issuance of the second tranche of the Series C redeemable convertible preferred stock in June 2026 will not be sufficient to fund its current business plan including related operating expenditure requirements through at least 12 months from the date of issuance of these financial statements. These conditions and events raise substantial doubt about the Company’s ability to continue as a going concern.
The Company’s long-term success is dependent upon its ability to successfully develop and market its pharmaceutical candidates, obtain additional capital when needed and, ultimately, to achieve profitable operations.
The Company plans to raise additional capital to continue funding its development efforts and operations. The Company is actively pursuing raising additional capital; however, there can be no assurance that funds will be available to the Company on acceptable terms, or on a timely basis, if at all based on both market conditions and management’s control. If the Company is unable to obtain an adequate level of capital needed to continue its research and development activities, the Company will need to delay, reduce or eliminate some or all of its planned activities and reduce costs. Doing so will likely have an adverse effect on the ability to execute the Company’s business plan. It is not considered probable that the Company’s plans to raise additional capital nor reduce discretionary spending will alleviate the substantial doubt regarding its ability to continue as a going concern.
The accompanying financial statements do not include any adjustments relating to the recoverability and classification of recorded asset amounts or the amounts and classification of liabilities that might result from the outcome of these uncertainties.
Reverse Stock Split
On September 10, 2026, the Company filed a certificate of amendment to the amended and restated certificate of incorporation to effect a 1-for-1.5002 reverse stock split of the Company’s outstanding shares of common stock and redeemable preferred stock. Upon the effectiveness of the reverse stock split, (i) all shares of outstanding common stock were adjusted; (ii) all shares of outstanding redeemable convertible preferred stock were adjusted; (iii) the number of shares of common stock for each outstanding option to purchase common stock was adjusted; and (iv) the
F-7
Table of Contents
exercise price of each outstanding option to purchase common stock was adjusted. All of the outstanding common stock share numbers (including shares of common stock subject to the Company’s options and as converted for the outstanding convertible preferred stock), share prices, exercise prices and per share amounts contained in the financial statements have been retroactively adjusted in the financial statements to reflect this reverse stock split for all periods presented. The par value per share and the authorized number of shares of common stock and preferred stock were not adjusted as a result of the reverse stock split.
Risks and Uncertainties
The Company is subject to risks and uncertainties common to early-stage companies in the biopharmaceutical industry, including, but not limited to, successful discovery and development of its product candidates, development by competitors of new technological innovations, dependence on key personnel, the ability to attract and retain qualified employees, protection of proprietary technology, compliance with governmental regulations, the change of internal trade relations, new legislation and tariffs, and the ability to secure additional capital to fund operations and commercial success of its product candidates. Ipsoprubart and any future product candidates the Company may develop will require extensive nonclinical and clinical testing and regulatory approval prior to commercialization. These efforts require significant amounts of additional capital, adequate personnel, and infrastructure and extensive compliance-reporting capabilities. Even if the Company’s product development efforts are successful, it is uncertain when, if ever, the Company will realize significant revenue from product sales.
2. Summary of Significant Accounting Policies
Basis of Presentation
The accompanying financial statements have been prepared in conformity with accounting principles generally accepted in the United States of America (“U.S. GAAP”), stated in U.S. dollars as of December 31, 2025 and 2024, and for the years then ended. Any reference in these notes to applicable guidance is meant to refer to U.S. GAAP as found in the Accounting Standards Codification (“ASC”) and Accounting Standards Updates (“ASU”) promulgated by the Financial Accounting Standards Board (“FASB”).
Use of Estimates
The preparation of financial statements requires management to make judgments, assumptions and estimates that affect the reported amounts of assets, liabilities, and expenses. Specific accounts that require management estimates include, but are not limited to, research and development accruals (including clinical trial accruals), the fair values of convertible preferred stock and common stock, and stock-based compensation. Management bases its estimates on historical experience and various other assumptions that are believed to be reasonable under the circumstances, the results of which form the basis for making judgments about the carrying value of assets and liabilities that are not readily apparent from other sources. Accordingly, actual results could differ materially from those estimates. To the extent that there are differences between management’s estimates and actual results, the Company’s future financial statement presentation, financial condition, results of operations, and cash flows may be affected.
Cash and Cash Equivalents
Cash and cash equivalents consist of highly liquid investments with original maturities of 90 days or less at the date of purchase, including funds held in government-backed money market accounts. Cash is deposited in checking accounts at financial institutions. Such deposits may, at times, exceed the limits established by the Federal Deposit Insurance Corporation (“FDIC”). The Company has not experienced any losses on its deposits.
Marketable Securities
The Company’s investments consist primarily of commercial paper, agency discount notes, corporate securities, and US government securities which are classified as available-for-sale. Available-for-sale investments are classified as current assets as they are highly liquid and available for use in current operations. The Company records investments at fair value with unrealized gains and losses recorded as a component of accumulated other comprehensive income on the balance sheets. Premiums and discounts are amortized (accreted) over the life of the related security as an adjustment to its yield. Dividend and interest income are recognized when earned. Realized gains and losses are included in other income (expense), net in the statements of operations and comprehensive loss and are derived using the specific identification method for determining the cost of investments sold.
F-8
Table of Contents
The Company assesses whether a loss on its investments has occurred due to declines in fair value or other market conditions. With respect to the Company’s debt securities, this assessment takes into account the severity and duration of the decline in value, its intent to sell the security, whether it is more likely than not that the Company will be required to sell the security before recovery of its amortized cost basis, and whether the Company expect to recover the entire amortized cost basis of the security (that is, whether a credit loss exists). A decline in the fair value of any debt security below cost that is deemed to be attributable to credit loss results in a charge to earnings and the corresponding establishment of an allowance for credit losses against the cost basis of the security. For the years ended December 31, 2025 and 2024, no credit losses were incurred.
Property and Equipment, Net
Property and equipment, net is stated at cost, less accumulated depreciation. Depreciation is calculated on a straight-line basis over the estimated useful lives of the related assets, generally ranging from three to five years. Maintenance and repair costs are expensed as incurred. Upon retirement or sale of the assets, the cost and related accumulated depreciation are removed from the balance sheet and the resulting gains or losses are recorded in the statements of operations and comprehensive loss.
Leases
The Company determines if an arrangement is a lease at inception. The Company classifies leases as either operating or finance leases. Operating lease right-of-use assets and lease liabilities are recognized at the commencement date based on the present value of lease payments over the lease term. Operating lease right-of-use assets also include any lease payments made at or before the commencement date, less any lease incentives received, and any initial direct costs incurred.
The Company uses its incremental borrowing rate (“IBR”) as the discount rate for leases when the rate implicit in the lease is not readily determinable. The IBR is estimated based on the rate of interest the Company would have to pay to borrow an amount equal to the lease payments on a collateralized basis over a similar term in a similar economic environment.
Operating lease cost is recognized on a straight-line basis over the lease term and is included in operating expenses in the statements of operations and comprehensive loss. Variable lease payments that do not depend on an index or rate are recognized as expense in the period in which the obligation is incurred. The Company accounts for lease components and non-lease components separately. Variable payments for the Company’s proportionate share of operating expenses and tax expenses are considered non-lease components and are not included in the measurement of the lease liability. For the Company’s long-term lease, the lease term includes the non-cancelable period of the lease plus any periods covered by an option to extend the lease when the Company is reasonably certain to exercise such option.
As of December 31, 2025, the Company did not have any short-term leases.
Impairment of Long-Lived Assets
Long-lived assets are reviewed annually for impairment or whenever events or changes in circumstances indicate that the carrying amount of an asset may not be recoverable. Factors that the Company considers in deciding when to perform an impairment review include significant underperformance of the business in relation to expectations, significant negative industry or economic trends and significant changes or planned changes in the use of the assets. If an impairment review is performed to evaluate a long-lived asset group for recoverability, the Company compares forecasts of undiscounted cash flows expected to result from the use and eventual disposition of the long-lived asset group to its carrying value. An impairment loss would be recognized when estimated undiscounted future cash flows expected to result from the use of an asset group are less than its carrying amount. The impairment loss would be based on the excess of the carrying value of the impaired asset group over its fair value, determined based on discounted cash flows. Long-lived assets consist of property and equipment and operating lease assets. There has been no impairment of long-lived assets for any of the periods presented.
Segment Information
ASC 280, Segment Reporting (“ASC 280”), defines operating segments as components of an enterprise where discrete financial information is available that is evaluated regularly by the chief operating decision-maker (“CODM”) in deciding how to allocate resources and in assessing performance. The Company operates as a single operating
F-9
Table of Contents
segment and has one reportable segment. The Company’s CODM is the chief executive officer, who has ultimate responsibility for the operating performance of the Company and the allocation of resources. The Company’s measure of segment profit or loss is net loss on a total company basis.
Concentrations of Credit Risk and Off-Balance Sheet Risk
Financial instruments that potentially subject the Company to significant concentrations of credit risk consist primarily of cash equivalents and marketable securities. The Company believes it is not exposed to any significant credit risk related to cash and cash equivalents because the Company places its investments in highly rated institutions.
The Company’s investment policy limits investments to certain types of securities issued by the U.S. government and its agencies, as well as institutions with investment-grade credit ratings and places restrictions on maturities and concentration by investment type and issuer. The Company is exposed to credit risk in the event of a default by issuers of marketable securities to the extent recorded on the balance sheets. As of December 31, 2025 and 2024, the Company had no off-balance sheet concentrations of credit risk.
Fair Value
Fair value is defined as the exchange price that would be received for an asset or paid to transfer a liability, or an exit price, in the principal or most advantageous market for that asset or liability in an orderly transaction between market participants on the measurement date.
Fair value is measured based on a three-level hierarchy of inputs, of which the first two are considered observable and the last unobservable. Unobservable inputs reflect the Company’s assumptions about current market conditions. The use of observable inputs is maximized, where available, and the use of unobservable inputs is minimized when measuring fair value. The three-level hierarchy of inputs is as follows:
Level 1 – Inputs are unadjusted quoted prices in active markets for identical assets or liabilities at the measurement date;
Level 2 – Inputs are unadjusted quoted prices in active markets for similar assets or liabilities, unadjusted quoted prices for identical or similar assets or liabilities in markets that are not active or other inputs that are observable or can be corroborated by observable market data for substantially the full term of the related assets or liabilities; and
Level 3 – Unobservable inputs that are significant to measuring the fair value of the assets or liabilities supported by little or no market data.
To the extent that the valuation is based on models or inputs that are less observable or unobservable in the market, determining fair value requires judgment. Accordingly, the degree of judgment exercised in determining fair value is greatest for instruments categorized in Level 3. A financial instrument’s level within the fair value hierarchy is based on the lowest level of any input that is significant to the fair value measurement.
The carrying amounts reflected in the balance sheet for cash, other receivables, prepaid expenses, accounts payable and accrued liabilities approximate their fair values due to the short-term nature of these assets and liabilities.
Research and Development Expenses
Research and development costs are expensed as incurred. Research and development expenses consist of costs incurred in connection with the development of the Company’s nonclinical and clinical product candidates. Research and development expenses consist of:
Direct program costs – including expenses incurred under agreements with consultants and third-party contract organizations that conduct research and development activities; costs related to the production of preclinical and clinical materials related to current or future product candidates paid to contract manufacturers; laboratory testing and other expenses related to the execution of nonclinical studies and clinical trials; and third-party license fees, including any milestone-based payments.
Unallocated costs – including personnel-related costs, such as salaries, benefits and stock-based compensation for employees engaged in research and development activities; and facility-related, costs not directly tied to a
F-10
Table of Contents
program, including rent, facility-related overhead, depreciation expense, expenses related to regulatory compliance and requirements, laboratory operations, information technology-related expenses, and other supplies and services.
Payments made prior to the receipt of goods and services to be used in research and development are deferred and recognized as expenses in the period in which the related goods are received or services are rendered. Certain prepaid amounts are classified as long-term based on contractual terms and conditions. In-licensing fees and other costs to acquire technologies used in research and development that have not yet received regulatory approval and that are not expected to have an alternative future use are expensed when incurred.
The Company has entered into agreements with outsourced contract manufacturing and development and clinical research vendors. The Company estimates accrued research and development expenses as of each balance sheet date based on facts and circumstances known at that time. The Company periodically confirms the accuracy of its estimates with internal management personnel and external service providers, and makes adjustments, if necessary. Research and development accruals are estimated based on the level of services performed, progress of the studies, including the phase or completion of events, and contracted costs. The estimated costs of research and development services provided, but not yet invoiced, are included in accrued expenses on the balance sheets. If the actual timing of the performance of services or the level of effort varies from the original estimates, the Company will adjust the accrual accordingly. Payments made under these arrangements in advance of the performance of the related services are recorded as prepaid expenses and other current assets until the services are rendered.
Stock-Based Compensation
In February 2022, the Company adopted the 2022 Stock Plan as an equity incentive program under which employees, non-employees and consultants of the Company may be offered the opportunity to acquire shares of the Company’s common stock. The Company’s stock-based compensation includes stock options issued under the 2022 Stock Plan and profits interest units which were granted under Electra Therapeutics LLC’s 2023 Equity Incentive Plan (the “Electra LLC Plan”).
The Company accounts for stock-based compensation in accordance with ASC 718, Compensation-Stock Compensation (“ASC 718”). The Company measures compensation expense for all stock-based payment awards for stock options and profits interest units granted to employees and non-employees based on the estimated fair values on the date of the grant which is then recorded straight-line over the requisite service period, generally the vesting period, for the entire award. Expense is adjusted for actual forfeitures of unvested awards as they occur. Fair values of the stock options were determined using the Black-Scholes option pricing model. Management determined the fair value of the awards with the assistance of third-party valuation specialists which determined the common stock value of the Company as an input into the Black-Scholes option pricing method.
Net Loss Per Share
Basic and diluted net loss per common share is presented in conformity with ASC 260, Earnings Per Share. The two-class method determines net loss per share for each class of common and participating securities according to dividends declared or accumulated and participation rights in undistributed earnings. The two-class method requires losses available to common stockholders for the period to be allocated between common and participating securities based upon their respective rights to share in the undistributed earnings as if all losses for the period had been distributed. The Company’s participating securities contractually entitled the holders of such shares to participate in dividends but did not contractually require the holders of such shares to participate in losses of the Company. Accordingly, in periods in which the Company reports a net loss attributable to common stockholders, such losses are not allocated to such participating securities.
Basic net loss per share is computed by dividing net loss attributed to common stockholders by the weighted-average number of common shares outstanding. Diluted loss per share is computed by giving effect to all potentially dilutive securities outstanding for the period using the treasury stock method or the if-converted method based on the nature of such securities.
For periods in which the Company reports net losses, diluted net loss per common share attributable to common stockholders is the same as basic net loss per common share attributable to common stockholders, because potentially dilutive common shares are not assumed to have been issued if their effect is anti-dilutive.
F-11
Table of Contents
Income Taxes
Income taxes are accounted for under the liability method. Deferred tax assets and liabilities are recognized based on the future tax consequences attributable to the differences between the financial statement carrying amounts of existing assets and liabilities and their respective tax bases and the operating loss and tax credit carryforwards. Valuation allowances are established when necessary to reduce deferred tax assets to the amount expected to be realized. Deferred tax assets and liabilities are measured at the balance sheet date using the enacted tax rates expected to apply to taxable income in the years when those temporary differences are expected to be recovered or settled. The effect on deferred tax assets and liabilities of a change in tax rates is recognized in the period such tax rate changes are enacted.
The Company records reserves for potential payments of taxes to various tax authorities related to uncertain tax positions. Amounts recognized are based on a determination of whether a tax benefit taken by the Company in its tax filings or positions is “more likely than not” to be sustained on audit. The amount recognized is equal to the largest amount that is more than 50% likely to be sustained. Interest and penalties associated with uncertain tax positions are recorded as a component of income tax expense.
Comprehensive Loss
Comprehensive loss includes net loss as well as other changes in stockholders’ deficit that result from transactions and economic events other than those with stockholders. The Company’s other comprehensive income (loss) is composed solely of unrealized gains or losses on marketable securities.
Emerging Growth Company Status
The Company is an emerging growth company, as defined in the Jumpstart Our Business Startups Act (“JOBS Act”). The JOBS Act provides that an “emerging growth company” can take advantage of an extended transition period for complying with new or revised accounting standards. The Company has elected not to “opt out” of such extended transition period, which means that when a standard is issued or revised and it has different application dates for public or private companies, the Company can adopt the new or revised standard at the time private companies adopt the new or revised standard and may do so until such time that the Company either (1) irrevocably elects to “opt out” of such extended transition period or (2) no longer qualifies as an emerging growth company. As a result, these financial statements may not be comparable to companies that comply with the new or revised accounting pronouncements as of public company effective dates.
Recently Adopted Accounting Pronouncements
In December 2023, the FASB issued a final standard on improvements to income tax disclosures, ASU 2023-09, Improvements to Income Tax Disclosures (“ASU 2023-09”). The standard requires disaggregated information about a reporting entity’s effective tax rate reconciliation as well as information on income taxes paid. The standard is intended to benefit investors by providing more detailed income tax disclosures that would be useful in making capital allocation decisions. This standard is effective for fiscal years beginning after December 15, 2024, and requires prospective application with the option to apply it retrospectively. The Company early adopted ASU 2023-09 on January 1, 2025 and applied the disclosure requirements on a retrospective basis. The early adoption did not have a material impact on the Company’s financial statements or disclosures.
In March 2024, the FASB issued ASU 2024-01, Compensation – Stock Compensation (Topic 718): Scope Application of Profits Interest and Similar Awards (“ASU 2024-01”), which provides illustrative guidance to help entities determine whether profits interest and similar awards should be accounted for as share-based payment arrangements under Topic 718. The standard is effective for fiscal years beginning after December 15, 2024 and interim periods within those fiscal years. The Company early adopted ASU 2024-01 on January 1, 2025. The early adoption of this standard did not have a material impact on the Company’s financial statements or disclosures.
Recently Issued Accounting Pronouncements
In November 2024, the FASB issued ASU 2024-03, Income Statement-Reporting Comprehensive Income-Expense Disaggregation Disclosures (Subtopic 220-40): Disaggregation of Income Statement Expenses (“ASU 2024-03”), requiring public entities to disclose additional information about specific expense categories in the notes to the financial statements on an interim and annual basis. ASU 2024-03 is effective for fiscal years beginning after December 15, 2026, and for interim periods beginning after December 15, 2027, with early adoption permitted. The Company is currently evaluating the impact of adopting ASU 2024-03.
F-12
Table of Contents
3. Related Party Transactions
Intercompany Services Agreements with Star Therapeutics, Inc.
Prior to January 1, 2024, the Company did not have any employees. The Company’s operations were conducted by employees of Star Therapeutics, Inc. (“Star Inc.”), a subsidiary of Star Therapeutics LLC (“Star LLC”). Star LLC was Electra’s parent company through June 2023. These related party transactions were governed by the Mutual Intercompany Services Agreements (“ISA”) between the Company and Star Inc., through its termination on October 31, 2025. Per the ISA, the Company bore all expenses incurred by Star Inc. in performing its duties in accordance with the agreements and in managing and operating the businesses of Electra with a minimal mark-up on expenses. Accordingly, expenses incurred and paid through the affiliated entity’s operating account on behalf of Electra were reimbursed by the Company to Star Inc. Operating expenses incurred by the affiliated entity included but were not limited to: service provider employee compensation expenses and other related overhead such as rent and general office expenses, utilities, information technology services, insurance, human resource management, taxes and other personnel costs.
For the years ended December 31, 2025 and 2024, the Company incurred expenses under the ISA totaling $1.1 million and $1.8 million, respectively, which were reimbursed by the Company to Star Inc. As of December 31, 2025 and 2024, the Company has outstanding amounts due to Star Inc. of $0 and $0.3 million, respectively. All outstanding accounts payable and accrued liabilities to Star Inc. were fully settled as of December 31, 2025.
Convertible Promissory Notes
On March 7, 2025, the Company issued convertible promissory notes with an aggregate principal amount of $20.0 million (the “Initial Notes”) to existing investors of the Company’s redeemable convertible preferred stock. On June 30, 2025, the Company issued additional convertible promissory notes with an aggregate principal amount of $20.0 million (the “Subsequent Notes” and, together with the Initial Notes, the “Notes”) to existing investors of the Company’s redeemable convertible preferred stock. The Notes bore interest at 8% per annum and were scheduled to mature on June 30, 2026. However, on October 15, 2025, the Company completed a qualified preferred stock equity financing through the issuance of Series C redeemable convertible preferred stock (the “Series C Preferred Stock”). In accordance with their original terms, the outstanding principal of the Notes and all accrued and unpaid interest, totaling $41.4 million, automatically converted into 4,080,265 shares of Series C Preferred Stock at a conversion price of $10.15 per share. The conversion price equaled the cash price per share paid by unrelated third-party investors participating in the Series C Preferred Stock financing. As of December 31, 2025, no convertible promissory notes remained outstanding (see Note 8 – Convertible Notes-Related Party and Note 10 – Redeemable Convertible Preferred Stock).
4. Fair Value Measurements
On a recurring annual basis, the Company measures certain financial assets and liabilities at fair value, including the Company’s cash equivalents and marketable securities. The following tables summarize the Company’s assets measured at fair value on a recurring basis, by level, within the fair value hierarchy as of December 31, 2025 and 2024 (in thousands):
| FAIR VALUE AS OF DECEMBER 31, 2025 | ||||||||||||||||
| LEVEL 1 | LEVEL 2 | LEVEL 3 | TOTAL | |||||||||||||
| Cash equivalents |
||||||||||||||||
| Money market funds |
$ | 26,517 | $ | — | $ | — | $ | 26,517 | ||||||||
| Agency discount notes |
— | 1,998 | — | 1,998 | ||||||||||||
| Commercial paper |
— | 3,995 | — | 3,995 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total cash equivalents |
26,517 | 5,993 | — | 32,510 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Marketable securities |
||||||||||||||||
| Agency discount notes |
— | 5,959 | — | 5,959 | ||||||||||||
| Commercial paper |
— | 29,559 | — | 29,559 | ||||||||||||
| Corporate securities |
— | 9,004 | — | 9,004 | ||||||||||||
| U.S. Government securities |
— | 8,988 | — | 8,988 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total marketable securities |
— | 53,510 | — | 53,510 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total assets measured at fair value |
$ | 26,517 | $ | 59,503 | $ | — | $ | 86,020 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
F-13
Table of Contents
| FAIR VALUE AS OF DECEMBER 31, 2024 | ||||||||||||||||
| LEVEL 1 | LEVEL 2 | LEVEL 3 | TOTAL | |||||||||||||
| Cash equivalents |
||||||||||||||||
| Money market funds |
$ | 10,019 | $ | — | $ | — | $ | 10,019 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total cash equivalents |
10,019 | — | — | 10,019 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Marketable securities |
||||||||||||||||
| Commercial paper |
— | 998 | — | 998 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total marketable securities |
— | 998 | — | 998 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total assets measured at fair value |
$ | 10,019 | $ | 998 | $ | — | $ | 11,017 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
Money market funds are included within Level 1 of the fair value hierarchy because they are valued using quoted market prices. Agency discount notes, commercial paper, corporate securities and U.S. government securities are classified within Level 2 of the fair value hierarchy as they take into consideration valuations obtained from third-party pricing services. The pricing services utilize industry standard valuation models, including both income-based and market-based approaches, for which all significant inputs are observable, either directly or indirectly, to estimate the fair value. These inputs include reported trades of and broker/dealer quotes on similar securities, issuer credit spreads, benchmark securities, prepayment/default projections based on historical data and other observable inputs. The Company does not currently have any Level 3 securities, and there were no transfers between Level 2 or 3 during 2025 or 2024. All debt securities are Level 2 and classified as available-for-sale.
As of December 31, 2025 and 2024, the Company had no financial liabilities that required fair value measurement.
5. Marketable Securities
The following tables summarize the amortized cost and the unrealized gains (losses) of the marketable securities presented within marketable securities and cash equivalents as of December 31, 2025 and 2024 (in thousands):
| AS OF DECEMBER 31, 2025 | ||||||||||||||||
| AMORTIZED COST |
NET UNREALIZED GAIN |
NET UNREALIZED (LOSS) |
ESTIMATED FAIR VALUE |
|||||||||||||
| Assets |
||||||||||||||||
| Agency discount notes |
$ | 7,955 | $ | 2 | $ | — | $ | 7,957 | ||||||||
| Commercial paper |
33,546 | 8 | — | 33,554 | ||||||||||||
| Corporate securities |
9,000 | 4 | — | 9,004 | ||||||||||||
| U.S. government securities |
8,982 | 6 | — | 8,988 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total marketable securities |
$ | 59,483 | $ | 20 | $ | — | $ | 59,503 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| AS OF DECEMBER 31, 2024 | ||||||||||||||||
| AMORTIZED COST |
NET UNREALIZED GAIN |
NET UNREALIZED (LOSS) |
ESTIMATED FAIR VALUE |
|||||||||||||
| Assets |
||||||||||||||||
| Commercial paper |
$ | 997 | $ | 1 | $ | — | $ | 998 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total marketable securities |
$ | 997 | $ | 1 | $ | — | $ | 998 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
F-14
Table of Contents
As of December 31, 2025, the weighted-average remaining maturity of the Company’s marketable securities was less than one year. The Company believes that an allowance for credit losses is unnecessary. As of December 31, 2025 and 2024, there were no securities in a continuous net unrealized loss position for more than 12 months. To date, the Company has not recorded any impairment charges on available-for-sale securities. Interest receivable as of December 31, 2025 and 2024 of $0.1 million and less than $0.1 million, respectively, are recorded as components of prepaid expenses and other current assets on the Company’s balance sheets. Additionally, accretion income due and accrued as of December 31, 2025 and 2024 of $0.1 million and less than $0.1 million, respectively, are recorded as components of the marketable securities balance.
6. Balance Sheet Components
Prepaids and Other Current Assets
Prepaids and other current assets consisted of the following as of December 31, 2025 and 2024 (in thousands):
| AS OF DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Prepaid clinical manufacturing |
$ | 8,516 | $ | 3,415 | ||||
| Prepaid insurance |
53 | — | ||||||
| Prepaid deposits and other |
179 | 165 | ||||||
|
|
|
|
|
|||||
| Total prepaid expenses |
$ | 8,748 | $ | 3,580 | ||||
|
|
|
|
|
|||||
Property and Equipment, Net
The Company acquired property and equipment during the year which was made up of lab equipment and office equipment for a total of $0.7 million. Total property and equipment as of December 31, 2025 and 2024 was $0.7 million and $0, respectively. Total accumulated depreciation as of December 31, 2025 and 2024 was $0.1 million and $0, respectively. For the years ended December 31, 2025 and 2024, the Company recorded $0.1 million, and $0 in depreciation expense, respectively, in the statements of operations and comprehensive loss.
Accrued Liabilities
Accrued liabilities consisted of the following as of December 31, 2025 and 2024 (in thousands):
| AS OF DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Accrued research and development |
$ | 429 | $ | 59 | ||||
| Accrued manufacturing |
566 | 369 | ||||||
| Accrued clinical trial costs |
1,706 | 547 | ||||||
| Accrued bonus |
2,014 | 1,259 | ||||||
| Accrued general and administrative |
330 | 304 | ||||||
|
|
|
|
|
|||||
| Total accrued liabilities |
$ | 5,045 | $ | 2,538 | ||||
|
|
|
|
|
|||||
7. Leases
Effective September 10, 2025 (the “Commencement Date”), the Company entered into a noncancelable operating lease agreement (the “Lease”) for approximately 14,055 rentable square feet of office and laboratory space located in South San Francisco, California (the “Premises”). The Lease has a three-year term expiring on September 30, 2028, and includes an initial rent abatement period. The Lease provides the Company with two consecutive options to extend the lease term for three years each, at the then-prevailing fair rental value; the Company has not included these extension options in the lease term as it is not reasonably certain to exercise either option. In addition to base rent, the Company is required to pay its proportionate share of the building’s operating and tax expenses, which are accounted for as variable lease payments and are not included in the measurement of the lease liability. The Company used an incremental borrowing rate of 6.45% to discount the lease payments, as the rate implicit in the Lease was not readily determinable.
F-15
Table of Contents
The components of lease expense were as follows as of December 31, 2025 (in thousands):
| AS OF DECEMBER 31, 2025 |
||||
| Operating lease expense (straight-line) |
$ | 260 | ||
| Variable lease expense (direct expenses) |
109 | |||
|
|
|
|||
| Total lease expense |
$ | 369 | ||
|
|
|
|||
Operating lease cost is recognized on a straight-line basis over the lease term. Cash paid for amounts included in the measurement of lease liabilities for the year ended December 31, 2025 was $0.3 million and was included in net cash used in operating activities in the Company’s statement of cash flows. Supplemental balance sheet information related to the Company’s operating lease was as follows as of December 31, 2025:
| AS OF DECEMBER 31, 2025 |
||||
| Weighted-average remaining lease term (years) |
2.75 | |||
| Weighted-average discount rate |
6.45 | % | ||
Future minimum lease payments under the Company’s non-cancelable operating lease were as follows as of December 31, 2025 (in thousands):
| YEAR ENDING DECEMBER 31, |
AMOUNT | |||
| 2026 |
$ | 629 | ||
| 2027 |
942 | |||
| 2028 |
725 | |||
|
|
|
|||
| Total undiscounted lease payments |
2,296 | |||
| Less: imputed interest |
(209 | ) | ||
|
|
|
|||
| Total operating lease liabilities |
$ | 2,087 | ||
|
|
|
|||
The future minimum lease payments above do not include variable lease payments for the Company’s proportionate share of building operating and tax expenses, which are recognized in the period incurred. The extension options are not included in the maturity analysis as the Company is not reasonably certain to exercise either option.
8. Convertible Notes – Related Party
On March 7, 2025, the Company issued Initial Notes to multiple investors. The Initial Notes bore interest at an annual rate of 8% per annum on the outstanding principal balance. Interest commenced on the closing date of March 7, 2025 and was computed on the basis of a year of 365 days for the actual number of days elapsed. The outstanding principal amount and all accrued and unpaid interest was due and payable upon demand by the Holders on or after June 30, 2026 (the “Maturity Date”). Each Holder committed to purchase additional notes in one or more future closings with the combined aggregate commitment of all Holders not to exceed $20.0 million.
On June 30, 2025, the Company issued the Subsequent Notes in the aggregate principal amount of $20.0 million pursuant to the commitment. The Subsequent Notes had terms identical to the Initial Notes, except for their respective principal amounts. The aggregate principal amount of the Notes outstanding was $40.0 million. The Notes were issued to existing investors of the Company’s redeemable convertible preferred stock, therefore, the issuances are considered related party transactions (see Note 3 – Related Party Transactions).
The Notes contained the following settlement provisions: (1) Upon the occurrence of a qualified preferred stock equity financing (a “Qualified Financing”) on or before the Maturity Date, the amount due automatically converts
F-16
Table of Contents
into fully paid and nonassessable shares of the preferred stock issued in such Qualified Financing at a price per share equal to the cash price per share paid by the other purchasers in the Qualified Financing; (2) if the Company completed a preferred stock equity financing that raised less than the Qualified Financing threshold of proceeds totaling $40.0 million, excluding note conversions, the Holders holding a majority of the principal outstanding under the Notes could elect to treat such financing as a Qualified Financing, with the amount due converting into preferred stock on the same terms, or hold the Notes until maturity; (3) upon the occurrence of a change of control, Holders were entitled to receive a cash payment equal to the amount due plus an applicable premium (the applicable premium was 20% if the change of control occurred prior to March 7, 2026, 50% if on or after March 7, 2026, and 100% premium if on or after the Maturity Date); and (4) upon an event of default, the amount due would become immediately due and payable in cash.
The Company elected the fair value option under ASC 825-10 to account for the Notes at the time of their initial recognition. Transaction costs incurred in connection with the issuance of the Notes were expensed as incurred. Interest expense from the Notes is presented in the statement of operations and comprehensive loss. For the year ended December 31, 2025, the Company recognized $1.4 million of expense in the statement of operations and comprehensive loss for the difference between the proceeds received and the fair value of the Notes, which was due to accrued interest on the Notes as of the conversion date. On issuance, total issuance costs of $0.1 million were expensed and recognized as general and administrative expense in the statement of operations and comprehensive loss.
On October 15, 2025, the Company completed a Qualified Financing through the issuance of Series C Preferred Stock (see Note 10 – Stockholders’ Deficit). Upon the closing of the Qualified Financing, the outstanding principal amount of the Notes and all accrued and unpaid interest thereon, totaling $41.4 million, were automatically converted into 4,080,265 shares of Series C Preferred Stock at a conversion price of $10.15 per share.
As of December 31, 2025, no convertible promissory notes remained outstanding. There was no gain or loss on extinguishment in the statement of operations and comprehensive loss for the year ended December 31, 2025.
9. Commitments and Contingencies
Commitments
The Company enters into contractual agreements with various suppliers in the normal course of its business. All contracts are terminable, with varying provisions regarding termination. If a contract with a specific vendor were to be terminated, the Company would only be obligated for the products or services that the Company had received through the time of termination.
In September 2025, the Company entered into a noncancelable operating lease agreement for approximately 14,055 rentable square feet of office and laboratory space in South San Francisco, California. The lease has a three-year term expiring on September 30, 2028, and includes an initial rent abatement period. The lease provides the Company with two consecutive options to extend the lease term for three years each, at the then-prevailing fair rental value. The Company has not included these extension options in the lease term as it is not reasonably certain to exercise either option. The lease provides for escalating annualized base rent payments starting at $0.9 million and increasing to $1.0 million in the final year of the lease. Remaining lease payments from January 1, 2026 through the end of the lease term total $2.3 million (see Note 7 – Leases).
The Company is also party to a mouse platform agreement with an unrelated third party. Under the agreement, the Company was granted non-exclusive licenses to use transgenic mice to generate antibodies and to exploit products containing the resulting antibodies (“Antibody Products”). Under the agreement, the Company paid a one-time initial technology access fee and is obligated to pay annual low five-digit maintenance fees per distinct antigen with which the Company immunized the mice, continuing until the first commercial sale of an Antibody Product. In addition, the agreement requires the Company to make certain development and commercial milestone payments upon the achievement of specified events by each Antibody Product, as well as pay royalties on net sales of Antibody Products. Aggregate milestone payments are up to the low double-digit millions of dollars per distinct Antibody Product. Royalties under the agreement are a low single digit percentage of net sales of all Antibody Products and are payable on a product-by-product and country-by-country basis beginning with the first commercial sale of the
F-17
Table of Contents
applicable Antibody Product in the applicable country and continuing for 10 years thereafter. The agreement remains in effect until the expiration of the last-to-expire royalty term, unless earlier terminated by the Company at will or by either party for the other party’s uncured material breach of a material provision of the agreement.
Contingencies
From time to time, the Company may have certain contingent liabilities that arise in the ordinary course of its business activities. The Company accrues a liability for such matters when it is probable that future expenditures will be made and that such expenditures can be reasonably estimated. Significant judgment is required to determine both probability and the estimated amount. There are no matters pending that the Company currently believes are reasonably possible or probable of having a material impact to the Company’s financial position, results of operations, or statements of cash flows.
In the normal course of operations, the Company may become involved in various legal proceedings. As of December 31, 2025 and 2024, the Company has not recorded accruals for probable losses related to any existing or pending litigation as the Company’s management has determined that there are no matters where a potential loss is probable and reasonably estimable. The Company does not believe that any existing or pending claims would have a material impact on the Company’s financial position, results of operations, or statements of cash flows.
Indemnification
In the normal course of business, the Company enters into contracts and agreements that contain a variety of representations and warranties and may provide for indemnification of the counterparty. The Company’s exposure under these agreements is unknown because it involves claims that may be made against it in the future but have not yet been made.
In accordance with the Company’s Amended and Restated Certificate of Incorporation (the “Certificate of Incorporation”) and Amended and Restated Bylaws, the Company has indemnification obligations to its officers and directors, subject to some limits, with respect to their service in such capacities. The Company has also entered into indemnification agreements with its directors and certain of its officers. To date, the Company has not been subject to any claims, and it maintains director and officer insurance that may enable it to recover a portion of any amounts paid for future potential claims.
The Company’s exposure under these agreements is unknown because it involves claims that may be made against it in the future but have not yet been made. The Company believes that the fair value of these indemnification obligations is minimal; accordingly, it has not recognized any liabilities relating to these obligations for any period presented.
10. Redeemable Convertible Preferred Stock
Although the Company’s redeemable convertible preferred stock is not mandatorily or currently redeemable, a liquidation or winding up of the Company could constitute an event outside its control. Therefore, all shares of redeemable convertible preferred stock have been presented outside of permanent equity due to being contingently redeemable. The redeemable convertible preferred stock is redeemable for cash or other assets upon the occurrence of a deemed liquidation event (including a change of control), which constitutes an event not solely within the Company’s control. Accordingly, the redeemable convertible preferred stock is classified outside of permanent equity and is presented as temporary equity in the balance sheets.
F-18
Table of Contents
The following table presents information related to redeemable convertible preferred stock as of December 31, 2025 (in thousands, except share and per share amounts):
| AS OF DECEMBER 31, 2025 | ||||||||||||||||||||
| SHARES AUTHORIZED |
SHARES ISSUED AND OUTSTANDING |
LIQUIDATION PREFERENCE |
CARRYING AMOUNT |
ORIGINAL ISSUE PRICE PER SHARE |
||||||||||||||||
| Series A |
22,601,626 | 15,065,741 | $ | 34,750 | $ | 34,650 | $ | 2.31 | ||||||||||||
| Series B |
9,309,492 | 6,205,494 | 84,019 | 83,836 | $ | 13.54 | ||||||||||||||
| Series C |
27,203,243 | 13,214,580 | 134,188 | 133,116 | $ | 10.15 | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||||||
| Total redeemable convertible preferred stock |
59,114,361 | 34,485,815 | $ | 252,957 | $ | 251,602 | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||||||
On October 15, 2025, pursuant to a Series C Preferred Stock Purchase Agreement (the “Purchase Agreement”), the Company issued 9,134,315 shares of its Series C Preferred Stock at a purchase price of $10.15 per share (the “Original Issue Price”) to multiple investors (the “Investors”) in a transaction referred to as the “Initial Closing”, resulting in gross proceeds of $92.8 million.
At the Initial Closing, previously issued Notes with aggregate principal and accrued interest of $41.4 million were automatically converted into 4,080,265 shares of Series C Preferred Stock at a conversion price equal to the Original Issue Price. $69.2 million of the $134.2 million Series C Preferred Stock was purchased by investors who had invested in the company prior to the Initial Closing, and are considered to be related parties.
The Purchase Agreement provides for a subsequent closing (the “Second Tranche Closing”) on or before the earlier of (1) October 1, 2026, or (2) the date set forth in a written election signed by the requisite investors (the “Second Tranche Requisite Investors” which means the Investors, other than defaulting investors, holding at least 60% of the then-outstanding shares, excluding any shares issued pursuant to the conversion of Notes), pursuant to which each Investor is obligated to purchase additional shares of Series C Preferred Stock at the Original Issue Price. Each Investor may elect to purchase some or all of its second tranche allocation early in one or more closings. If an investor defaults on its Second Tranche purchase obligation (a “Defaulting Investor”), all shares of the Company’s Series B redeemable convertible preferred stock (the “Series B Preferred Stock”) and Series C Preferred Stock held by such Defaulting Investor and its affiliates (the “Subject Shares”) shall automatically convert into Common Stock at a conversion price equal to one-tenth of the applicable conversion price in effect at the time of conversion (the “Special Mandatory Conversion”). The Series C Preferred Stock and the associated rights for the Second Tranche Closing were contractually linked and unable to be sold or exercised separately except in limited affiliate transfer scenarios. As such, the Second Tranche Closing was defined as an embedded feature within the Series C Preferred Stock financing rather than a freestanding instrument, and no liability was recorded for the second tranche closing in 2025.
Dividend: Each holder of redeemable convertible preferred stock is entitled to receive non-cumulative dividends at the rate of 8% per annum for each share of redeemable convertible preferred stock outstanding, when, as and if declared by the Company’s Board of Directors (the “Board of Directors”). These dividends are payable in preference to common stock dividends. To date, the Company has not declared or paid any dividends.
Liquidation: In the event of any liquidation event, either voluntary or involuntary, the holders of preferred stock shall be entitled to receive out of the proceeds or assets available for distribution, prior and in preference to any distributions to the holders of common stock, an amount per share equal to the sum of the applicable original issuance price, plus declared but unpaid dividends on such share. If the proceeds distributed among the holders of the preferred stock are insufficient to permit the payment to such holders, then the entire proceeds available for distribution shall be distributed ratably among the holders of preferred stock in the same proportion to respective amounts, which would be payable if paid in full.
Conversion for Series A and B Preferred Stock: Each share of preferred stock shall be convertible, at the option of the holder, at any time after the date of issuance of such share into such number of shares of common stock determined by dividing the original issue price for preferred stock series by the conversion price. The conversion price is the same as the original issuance price for Series A Preferred Stock and Series B Preferred Stock.
F-19
Table of Contents
Conversion of each series of redeemable convertible preferred stock into common stock is automatic upon the earlier of: (1) the closing of the sale of common stock in a firm commitment underwritten public offering that results in at least $100 million of gross proceeds at a minimum price per share of $10.15, subject to adjustment in the event of a stock split; or (2) the date, or the occurrence of an event, specified by vote or written consent or agreement of the Requisite Investors (“Requisite Investors” means Investors holding at least 70% of the shares of Series B Preferred Stock and Series C Preferred Stock then outstanding (voting together as a single class and not as separate series, and on an as-converted basis).
Conversion for Series C Preferred Stock: Each share of preferred stock shall be convertible, at the option of the holder, at the earlier of: (1) the day after the Second Tranche Closing; (2) a Liquidation Event; (3) the termination of the Second Tranche obligation; or (4) another date designated by the Company and the Requisite Investors, excluding any shares held by defaulting investors.
The conversion price is the same as the original issuance price for Series C Preferred Stock.
Conversion of each series of redeemable convertible preferred stock into common stock is automatic upon the earlier of: (1) the closing of the sale of common stock in a firm commitment underwritten public offering that results in at least $100.0 million of gross proceeds at a minimum price per share of $10.15, subject to adjustment in the event of a stock split; or (2) the date, or the occurrence of an event, specified by vote or written consent or agreement of the Requisite Investors.
The conversion price is also subject to standard anti-dilutive adjustments for stock splits, combinations, dividends, distributions, reclassifications, and recapitalizations, as set forth in the Certificate of Incorporation. Any downward adjustment to the conversion price of the Series C Preferred Stock may be waived, prospectively or retroactively, by the consent or vote of holders of at least 60% of the then-outstanding shares of Series C Preferred Stock (voting as a single class on an as-converted basis).
Voting Rights: The holder of each share of preferred stock shall have the right to one vote for each share of common stock into which shares of preferred stock could then be converted. The Board of Directors of the Company is set at 7 members with 1 vacancy, appointed as follows: 2 directors from the Holders of Series B Preferred Stock, 1 director from the Holders of Series C Preferred Stock, 4 directors from the Holders of a majority of outstanding capital stock (voting together as a single class on an as-converted basis).
As of December 31, 2025, a deemed liquidation event was not considered probable of occurring. Accordingly, the redeemable convertible preferred stock has not been remeasured to its redemption value. The Company will reassess the probability of a deemed liquidation event and the classification and measurement of the redeemable convertible preferred stock at each reporting period end.
11. Stockholders’ Deficit
Stock-Based Compensation
The Company recognized stock-based compensation in the years ended December 31, 2025 and 2024 as follows (in thousands):
| YEAR ENDED DECEMBER 31, |
||||||||
| 2025 | 2024 | |||||||
| Research and development expense – PI Units |
$ | 123 | $ | 258 | ||||
| Research and development expense – Options |
746 | 729 | ||||||
|
|
|
|
|
|||||
| Total research and development expense |
869 | 987 | ||||||
| General and administrative expense – PI Units |
357 | 795 | ||||||
| General and administrative expense – Options |
548 | 527 | ||||||
|
|
|
|
|
|||||
| Total general and administrative expense |
905 | 1,322 | ||||||
|
|
|
|
|
|||||
| Total stock-based compensation expense |
$ | 1,774 | $ | 2,309 | ||||
|
|
|
|
|
|||||
F-20
Table of Contents
Profits Interest Units
In December 2018, Star LLC adopted the 2018 Equity Incentive Plan (the “Star LLC Plan”) pursuant to which it could grant profits interest unit awards to its employees, managers, and consultants, including those hired and engaged by its affiliated entities. In June 2023, Star LLC’s members formed Electra Therapeutics LLC (“Electra LLC”) and contributed their Series A redeemable convertible preferred shares to Electra LLC in exchange for the Series B preferred units, Series A preferred units, Series Seed preferred units, and common units of Electra LLC. This transaction transferred all of Star LLC’s equity ownership in the Company to Electra LLC, a sister company of Star LLC immediately after the spin-off. In connection with the spin-off, Electra LLC adopted the 2023 Equity Incentive Plan (the “Electra LLC Plan”) and granted profits interest units (“PI Units”) to Star LLC’s profits interest unit holders who supported the operations of Electra through the ISA. On December 31, 2023, Star LLC terminated 16 employees who were immediately employed by Electra, effective January 1, 2024. On January 1, 2024, the 4.5 million PI Units previously awarded to these holders for services rendered under the MISA were remeasured with a weighted average fair value of $1.04 per PI Unit. All other terms of the PI Units remained the same. The total value of the PI Units under the Electra LLC Plan that vested during the years ended December 31, 2025 and 2024 was recorded by the Company as stock-based compensation expense in the amount of $0.3 million and $1.0 million, respectively. Under the provisions of the Electra LLC Plan, the board of managers of Electra LLC (the “Board of Managers”) may grant PI Units to its employees, managers, and consultants, including those hired and engaged by its affiliated entities (collectively, the “Participants”). PI Units are common units issued to Participants with a threshold amount. In the event of distribution by Electra LLC, the proceeds distributed to the holder would be reduced by the threshold amount. PI Units are economically similar to a stock option award and vest based on time or performance-based milestones, as determined by the Board of Managers of Electra LLC and stipulated in the grant agreements.
Profits interest units generally vest 25% after one year, with the remainder vesting monthly over the following three-year period. The underlying terms and intended purpose of the PI Units are more akin to stock-based compensation for employees and non-employees than a performance bonus or profit-sharing arrangement.
Determination of Fair Value
The estimated grant-date fair value of the PI Units was calculated using the Black-Scholes option pricing model, based on the following assumptions:
Fair Value – The fair value of Electra LLC was determined on January 1, 2024 (modification date), and thereafter when there has been a grant by the Board of Managers with the assistance of third-party valuation specialists. Because there has been no public market for the Company’s common units to determine the enterprise value, the Board of Managers exercised reasonable judgment and considered a number of objective and subjective factors to determine the best estimate of the fair market value, which included important developments in the Company’s operations, the prices at which Electra LLC sold its convertible preferred units, the rights, preferences and privileges of Electra LLC’s convertible preferred units relative to those of Electra LLC’s common units, actual operating results, financial performance, external market conditions in the life sciences industry, general U.S. market conditions, equity valuations of comparable public companies, and the lack of marketability of Electra LLC’s common units.
Expected Term – The expected term was estimated based on management’s expectations for a future liquidity event of Electra LLC.
Expected Volatility – The Company had limited information on the volatility of PI Units as the units were not actively traded on any public markets. The expected volatility was derived from the historical unit volatilities of comparable peer public companies.
Risk-Free Interest Rate – The risk-free interest rate was based on the U.S. Treasury yield curve in effect at the date of grant for zero-coupon U.S. Treasury notes with maturities approximately equal to the PI Units’ and unit options expected term.
Discount for Lack of Marketability – Since the PI Units are interests in a closely held entity, the Company also considered appropriate adjustments to recognize the lack of marketability inherently present in interests of this type. The concluded adjustments for lack of marketability were supported by put option analyses.
F-21
Table of Contents
No PI Units were granted during the year ended December 31, 2025. There were 1,211,639 PI Units granted in 2024 which have a threshold price of $1.77 per share and a weighted-average grant date fair value of $0.83 per share. The table below summarizes the assumptions used to determine the grant-date fair value of the PI Units granted in the year ended December 31, 2024.
| YEAR ENDED DECEMBER 31, 2024 |
||||
| Expected term |
1.52 years | |||
| Expected volatility |
90.0 | % | ||
| Risk-free interest rate |
4.93 | % | ||
| Discount for lack of marketability |
34.5 | % | ||
Stock Incentive Plan
In 2022, the Board of Directors approved the adoption of the 2022 Stock Plan (the “Plan”). The Plan allows for the award of incentive and non-qualified stock options and restricted stock to employees, outside directors, and consultants of the Company. A total of 5,663,057 and 2,363,471 shares of Common Stock were reserved for issuance under the Plan as of December 31, 2025 and 2024, respectively. As of December 31, 2025, 3,367,661 shares were available for award under the Plan. The Board of Directors administers the Plan and determines the exercise price of options, purchase price for restricted stock, the rates at which awards vest, and the other terms and conditions of the awards. Options and restricted stock generally vest 25% upon the first anniversary of the grant date and an additional 1/48th of the shares vest each month of continuous service thereafter for 36 months, and expire 10 years from the date of grant. Stock options are eligible for early exercise. No tax benefits were realized from options and other share-based payment arrangements during the year.
The Company grants stock options at exercise prices deemed by the Board of Directors to be equal to the fair value of the common stock at the time of grant. The fair value of common stock has been determined by the Board of Directors at each stock option measurement date based on a variety of different factors, including contemporaneous independent valuations performed by an independent third-party valuation firm; the prices per share of redeemable convertible preferred stock sold to investors in arm’s length transactions, and the rights, preferences, and privileges of the redeemable convertible preferred stock relative to the Company’s common stock; the Company’s stage of development and material risks related to the business; results of operations and financial position, including levels of available capital resources; progress of research and development activities; the lack of marketability of the Company’s common stock as a private company; the status of strategic transactions; the hiring of key personnel and the experience of management; the likelihood and timing of achieving a liquidity event, such as an initial public offering or a sale of the Company, given prevailing market conditions; market performance of comparable publicly traded companies; trends and developments in the life sciences and biopharmaceutical industry; and external market conditions affecting the industry sector.
During the years ended December 31, 2025 and 2024, the Company calculated the value of the stock options using the Black-Scholes option pricing model with the following assumptions:
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Expected term (in years) |
5.70 - 6.08 | 6.08 | ||||||
| Expected volatility |
93.1% - 103.4 | % | 94.9% - 103.5 | % | ||||
| Expected dividend yield |
— | — | ||||||
| Risk-free interest rate |
3.8% - 4.4 | % | 3.9% - 4.5 | % | ||||
The Black-Scholes option pricing model requires the use of subjective assumptions to determine the fair value of stock-based awards, including:
| ∎ | Expected Term – using the simplified method which calculates the expected term as the average of the time-to-vesting and the contractual life of the stock options. |
F-22
Table of Contents
| ∎ | Expected Volatility – based on a study of publicly traded industry peer companies which were identified based on industry, stage of development, size, and financial leverage of potential comparable companies. Historical volatility is calculated based on a period of time commensurate with the expected term assumption. |
| ∎ | Expected Dividend Yield – assuming the expected dividend yield is zero, as the Company has no history or expectation of paying cash dividends on common stock in the foreseeable future. |
| ∎ | Risk-Free Interest Rate – based on the yield available on U.S. Treasury zero-coupon issues similar in duration to the expected term of the award. |
The Company records stock-based compensation expense on options as they vest and records forfeitures as they occur.
The option activity of the Plan for the year ended December 31, 2025 was as follows:
| STOCK OPTION ACTIVITY |
OPTIONS OUTSTANDING | |||||||||||||||
| NUMBER OF OPTIONS |
WEIGHTED- AVERAGE EXERCISE PRICE PER SHARE |
WEIGHTED- AVERAGE REMAINING CONTRACTUAL TERM (IN YEARS) |
AGGREGATE INTRINSIC VALUE (IN THOUSANDS) |
|||||||||||||
| Balances as of December 31, 2024 |
1,826,821 | $ | 4.84 | 8.47 | $ | 493 | ||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Options granted |
560,490 | $ | 4.96 | — | ||||||||||||
| Options forfeited |
(75,583 | ) | $ | 5.05 | — | |||||||||||
| Options cancelled |
(45,661 | ) | $ | 5.21 | — | |||||||||||
|
|
|
|||||||||||||||
| Balances as of December 31, 2025 |
2,266,067 | $ | 4.86 | 8.05 | $ | 423 | ||||||||||
|
|
|
|||||||||||||||
| Exercisable as of December 31, 2025 |
1,090,694 | $ | 4.90 | 7.08 | $ | 224 | ||||||||||
| Vested and expected to vest as of December 31, 2025 |
2,266,067 | $ | 4.86 | 8.05 | $ | 423 | ||||||||||
As of December 31, 2025 the total gross unrecognized stock-based compensation expense related to unvested stock options aggregated to $4.2 million. These expenses are expected to be recognized over a weighted-average period of 3.04 years. During the years ended December 31, 2025 and 2024, the Company granted 560,490 and 478,831 options, respectively, with a weighted-average grant date fair value of $4.01 and $3.98 per share, respectively. The total fair value of options vested during the years ended December 31, 2025 and 2024 was $1.7 million and $1.2 million, respectively. The aggregate intrinsic values of options outstanding, exercisable, vested and expected to vest were calculated as the difference between the exercise price of the options and fair value of the Company’s common stock as of December 31, 2025.
12. Income Taxes
As the Company was in a net loss position for the years ended December 31, 2025 and 2024, no income tax expense was recorded.
The components of the Income (loss) before provision for income taxes were as follows for the years ended December 31, 2025 and 2024 (in thousands):
| YEAR ENDED DECEMBER 31, |
||||||||
| 2025 | 2024 | |||||||
| U.S. |
(61,975 | ) | (24,786 | ) | ||||
| Foreign |
— | — | ||||||
|
|
|
|
|
|||||
| Loss before provision for income taxes |
$ | (61,975 | ) | $ | (24,786 | ) | ||
|
|
|
|
|
|||||
F-23
Table of Contents
The components of the Company’s deferred tax asset (liabilities) as of December 31, 2025 and 2024 were as follows (in thousands):
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Deferred tax assets: |
||||||||
| Net operating loss carryforward |
$ | 22,601 | $ | 13,265 | ||||
| Research and development costs |
11,902 | 9,049 | ||||||
| Credit carryforwards |
491 | 491 | ||||||
| Accruals and reserves |
424 | 265 | ||||||
| Fixed assets |
5 | — | ||||||
| Intangible assets |
43 | 49 | ||||||
| Stock-based compensation |
596 | 512 | ||||||
| Lease liability |
439 | — | ||||||
|
|
|
|
|
|||||
| Total deferred tax assets |
36,501 | 23,631 | ||||||
| Deferred tax liabilities: |
||||||||
| Right-of-use asset |
(444 | ) | — | |||||
|
|
|
|
|
|||||
| Total net deferred tax assets |
36,057 | 23,631 | ||||||
| Less: valuation allowance |
(36,057 | ) | (23,631 | ) | ||||
|
|
|
|
|
|||||
| Deferred tax asset, net |
$ | — | $ | — | ||||
|
|
|
|
|
|||||
The following table provides a reconciliation of the U.S. statutory income tax rate to the Company’s provision for income taxes and respective effective tax rate disaggregated by required category for the years ended December 31, 2025 and 2024 in accordance with ASU 2023-09.
| YEAR ENDED DECEMBER 31, | ||||||||||||||||
| 2025 | 2024 | |||||||||||||||
| RATE | AMOUNT | RATE | AMOUNT | |||||||||||||
| Income tax expense (benefit) at statutory federal rate |
21.0% | (13,014 | ) | 21.0% | (5,187 | ) | ||||||||||
| Tax credits |
||||||||||||||||
| Federal research and development credits |
0.0% | — | 2.6% | (644 | ) | |||||||||||
| Changes in valuation allowance |
(20.0% | ) | 12,412 | (18.7% | ) | 4,615 | ||||||||||
| Nontaxable or nondeductible items |
||||||||||||||||
| Convertible note interest |
(0.5% | ) | 301 | 0 | % | — | ||||||||||
| Stock-based compensation |
(0.4% | ) | 288 | (2.2 | %) | 551 | ||||||||||
| Other permanent items |
(0.1% | ) | 30 | (0.1 | %) | 21 | ||||||||||
| Changes in unrecognized tax benefits |
||||||||||||||||
| Federal research and development credits |
0.0% | — | (2.6 | %) | 644 | |||||||||||
| Other reconciling items |
||||||||||||||||
| Other |
0.0% | (17 | ) | 0.0 | % | — | ||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Effective income tax rate |
0.0% | — | 0.0 | % | — | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
A valuation allowance is provided for deferred tax assets where the recoverability of the assets is uncertain. The determination to provide a valuation allowance is dependent upon the assessment of whether it is more likely than not that sufficient future taxable income will be generated to utilize the deferred tax assets. Based on the weight of the available evidence, which includes the Company’s historical operating losses, the Company provided a full valuation allowance against the deferred tax assets resulting from the tax losses and credits carried forward. The ultimate tax benefit will be realized upon the achievement of continued sustained future operating profits and deriving taxable income.
F-24
Table of Contents
On July 4, 2025, President Trump signed the tax act referred to as The One Big Beautiful Bill (“OBBB”) Act, which includes comprehensive U.S. corporate tax legislation. The legislation includes the permanent extension and modification of provisions originally introduced under the Tax Cuts and Jobs Act of 2017 and the introduction of new provisions. Key provisions include the permanent restoration of bonus depreciation allowances, changes to the limitations on deductibility of business interest expense, and the reintroduction of immediate expensing of U.S. domestic research and development costs. The impact on current and deferred taxes for tax law changes is reported in continuing operations in the interim period that includes the enactment date. The Company has determined that the new act would not have a material impact to its tax provision due to full valuation allowance. The Company will continue to assess the tax accounting impacts, as well as the various state legislation and conformity rules, as more information becomes available.
During the year ended December 31, 2025, the Company capitalized $23.7 million of foreign research and development. In addition, during the year ended December 31, 2025, the Company recorded $10.4 million of amortization expenses related to previously capitalized domestic and foreign research and development expenditures.
As of December 31, 2025, the Company had net operating loss carryforwards of $102.6 million for federal income taxes and $14.6 million for state income taxes. If not utilized, the Federal net operating losses will not expire and the state net operating losses will begin to expire in 2038. As of December 31, 2025, the Company had research and development credit carryforwards of $0.5 million and less than $0.1 million for federal and California income taxes, net of reserves, respectively. If not utilized, the federal carryforwards will begin to expire in various amounts beginning in 2041. The state tax credit can be carried forward indefinitely.
Utilization of the net operating loss and tax credit carryforwards may be subject to a substantial annual limitation due to limitations from a change in ownership as provided by Section 382 of the Internal Revenue Code of 1986, as amended (“IRC”), and similar state provisions. The annual limitation may result in the expiration of net operating losses and tax credits before utilization. In the event that the Company had a change of ownership, utilization of the net operating loss and tax credit carryforwards may be restricted. The Company performed a Section 382 study through December 31, 2025, and concluded that the Company did not undergo an ownership change within the meaning of Section 382.
All of the Company’s tax returns remain open to examination. The resolution of the uncertain tax position would not affect the income tax rate due to the full valuation allowance on deferred tax assets.
The unrecognized tax benefit is related to the Company’s reserves on Federal and California research and development tax credits. For the years ended December 31, 2025 and 2024, the activity related to the unrecognized tax benefits is as follows (in thousands):
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Balance at beginning of year |
$ | 1,442 | $ | 798 | ||||
| Changes related to prior tax positions |
— | 644 | ||||||
| Increases related to current tax positions |
— | — | ||||||
|
|
|
|
|
|||||
| Balance at end of year |
$ | 1,442 | $ | 1,442 | ||||
|
|
|
|
|
|||||
13. Segments
The Company’s CODM is its Chief Executive Officer, who reviews financial information presented on a net loss basis as reported in the statement of operations in order to make decisions about allocating resources and assessing performance for the entire Company. The CODM also utilizes the Company’s annual budget and strategic plan, as a key input to resource allocation. The CODM function approves of key operating and strategic decisions. The CODM function views the Company’s operations and manages its business as a single reportable operating segment.
The Company monitors its cash, cash equivalents and investments as reported on the Company’s balance sheets to determine funding needed for furthering its research and development activities, while continuing general and
F-25
Table of Contents
administrative operations. The CODM function is regularly provided with the following significant segment expenses. Significant expenses include research and development and general and administrative expenses, which are each separately presented in the Company’s statements of operations. The CODM reviews significant expenses within both the research and development and the general and administrative categories. Other segment items within net loss include interest income, interest expense, and other income (expense). No product revenue has been generated since inception and all assets are held in the United States. See the financial statements for other financial information regarding the Company’s operating segment. The CODM is regularly provided with the following significant segment expenses:
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Operating expenses: |
||||||||
| Research and development: |
||||||||
| Direct program costs: |
||||||||
| Ipsoprubart |
$ | 26,100 | $ | 6,794 | ||||
| Other programs |
12,594 | 4,360 | ||||||
| Unallocated costs: |
||||||||
| Personnel-related |
9,339 | 6,115 | ||||||
| Facilities, overhead and other |
2,354 | 1,998 | ||||||
|
|
|
|
|
|||||
| Total research and development |
50,387 | 19,267 | ||||||
| General and administrative: |
||||||||
| Personnel-related |
3,370 | 2,834 | ||||||
| Third-party service providers |
7,178 | 3,628 | ||||||
| Facilities, overhead and other |
954 | 588 | ||||||
|
|
|
|
|
|||||
| Total general and administrative |
11,502 | 7,050 | ||||||
| Total operating expenses |
61,889 | 26,317 | ||||||
|
|
|
|
|
|||||
| Loss from operations |
(61,889 | ) | (26,317 | ) | ||||
|
|
|
|
|
|||||
| Other income (expense), net: |
||||||||
| Interest income |
1,362 | 1,524 | ||||||
| Interest expense – related party |
(1,433 | ) | — | |||||
| Other income (expense) |
(15 | ) | 7 | |||||
|
|
|
|
|
|||||
| Total other income (expense), net: |
(86 | ) | 1,531 | |||||
|
|
|
|
|
|||||
| Net loss |
$ | (61,975 | ) | $ | (24,786 | ) | ||
|
|
|
|
|
|||||
14. Net Loss Per Share Attributable to Common Stockholders
The following table sets forth the computation of basic and diluted net loss per share attributable to common stockholders (in thousands, except share and per share data):
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Numerator: |
||||||||
| Net loss |
$ | (61,975 | ) | $ | (24,786 | ) | ||
|
|
|
|
|
|||||
| Denominator: |
||||||||
| Weighted-average shares outstanding in computing net loss per share, basic and diluted |
29,329 | 29,329 | ||||||
|
|
|
|
|
|||||
| Net loss per share attributable to common stockholders, basic and diluted |
$ | (2,113.10 | ) | $ | (845.10 | ) | ||
|
|
|
|
|
|||||
F-26
Table of Contents
The following outstanding shares of potentially dilutive securities were excluded from the computation of diluted net loss per share for the periods presented because including them would have been anti-dilutive:
| YEAR ENDED DECEMBER 31, | ||||||||
| 2025 | 2024 | |||||||
| Redeemable convertible preferred stock |
34,485,815 | 21,271,235 | ||||||
| Stock options |
2,266,067 | 1,826,821 | ||||||
|
|
|
|
|
|||||
| Total |
36,751,882 | 23,098,056 | ||||||
|
|
|
|
|
|||||
Electra LLC PI Units are excluded as they are not potentially dilutive securities to the Company.
15. Subsequent events
The Company evaluated subsequent events through June 26, 2026, the date on which these financial statements were available for issuance. No subsequent events have been identified for disclosure to or adjustment in the financial statements, other than the matter noted below.
On June 18, 2026, pursuant to the Series C Preferred Stock Purchase Agreement, the Company closed on the Second Tranche Closing and issued shares of its Series C Preferred Stock at the Original Issue Price of $10.15 per share on the same terms as the Initial Tranche. The Second Tranche Closing was made up of the same investors as those who participated in the Initial Tranche in October 2025 and were committed to the Second Tranche. The Second Tranche Closing resulted in the issuance of 4,918,477 shares of Series C Preferred Stock for total gross proceeds of $49.9 million.
F-27
Table of Contents
Condensed Balance Sheets
(Unaudited)
(In thousands, except share and per share amounts)
| JUNE 30, 2026 |
DECEMBER 31, 2025 |
|||||||
| Assets |
||||||||
| Current assets: |
||||||||
| Cash and cash equivalents |
$ | 66,507 | $ | 33,008 | ||||
| Marketable securities |
31,218 | 53,510 | ||||||
| Other receivables |
264 | 220 | ||||||
| Prepaid expenses and other current assets |
2,372 | 8,748 | ||||||
|
|
|
|
|
|||||
| Total current assets |
100,361 | 95,486 | ||||||
| Property and equipment, net |
579 | 674 | ||||||
| Right-of-use assets |
1,830 | 2,110 | ||||||
| Other assets, non-current |
3,728 | 1,098 | ||||||
|
|
|
|
|
|||||
| Total assets |
$ | 106,498 | $ | 99,368 | ||||
|
|
|
|
|
|||||
| Liabilities, redeemable convertible preferred stock, and stockholders’ deficit |
||||||||
| Current liabilities: |
||||||||
| Accounts payable |
$ | 4,606 | $ | 4,076 | ||||
| Accrued liabilities |
8,193 | 5,045 | ||||||
| Operating lease liability |
827 | 605 | ||||||
|
|
|
|
|
|||||
| Total current liabilities |
13,626 | 9,726 | ||||||
| Operating lease liability, net of current portion |
1,156 | 1,482 | ||||||
|
|
|
|
|
|||||
| Total liabilities |
14,782 | 11,208 | ||||||
|
|
|
|
|
|||||
| Commitments and contingencies (Note 9) |
||||||||
| Series A redeemable convertible preferred stock: $0.0001 par value – 22,601,626 shares authorized, 15,065,741 issued and outstanding as of June 30, 2026 and December 31, 2025 (liquidation preference of $34,750 as of June 30, 2026) |
34,650 | 34,650 | ||||||
| Series B redeemable convertible preferred stock: $0.0001 par value – 9,309,492 shares authorized, 6,205,494 issued and outstanding as of June 30, 2026 and December 31, 2025 (liquidation preference of $84,019 as of June 30, 2026) |
83,836 | 83,836 | ||||||
| Series C redeemable convertible preferred stock: $0.001 par value – 27,203,243 shares authorized as of June 30, 2026 and December 31, 2025. 18,133,057 and 13,214,580 shares issued and outstanding as of June 30, 2026 and December 31, 2025, respectively (liquidation preference of $184,133 as of June 30, 2026) |
192,685 | 133,116 | ||||||
| Stockholders’ deficit: |
||||||||
| Common stock: $0.0001 par value – 71,000,000 authorized shares as of June 30, 2026 and December 31, 2025; 29,329 shares issued and outstanding as of June 30, 2026 and December 31, 2025 |
— | — | ||||||
| Additional paid-in-capital |
2,448 | 9,515 | ||||||
| Accumulated deficit |
(221,890 | ) | (172,978 | ) | ||||
| Accumulated other comprehensive income (loss) |
(13 | ) | 21 | |||||
|
|
|
|
|
|||||
| Total stockholders’ deficit |
(219,455 | ) | (163,442 | ) | ||||
|
|
|
|
|
|||||
| Total liabilities, redeemable convertible preferred stock and stockholders’ deficit |
$ | 106,498 | $ | 99,368 | ||||
|
|
|
|
|
|||||
See accompanying notes to the condensed financial statements.
F-28
Table of Contents
ELECTRA THERAPEUTICS, INC.
Condensed Statements of Operations
(Unaudited)
(In thousands except share and per share amounts)
| SIX MONTHS ENDED JUNE 30, |
||||||||
| 2026 | 2025 | |||||||
| Operating expenses: |
||||||||
| Research and development |
$ | 41,637 | $ | 18,792 | ||||
| General and administrative |
8,593 | 5,921 | ||||||
| General and administrative – related party |
— | 656 | ||||||
|
|
|
|
|
|||||
| Total operating expenses |
50,230 | 25,369 | ||||||
|
|
|
|
|
|||||
| Loss from operations |
(50,230 | ) | (25,369 | ) | ||||
|
|
|
|
|
|||||
| Other income (expense), net: |
||||||||
| Interest income |
1,320 | 337 | ||||||
| Interest expense – related party |
— | (327 | ) | |||||
| Other income (expense) |
(2 | ) | (2 | ) | ||||
|
|
|
|
|
|||||
| Total other income (expense), net |
1,318 | 8 | ||||||
|
|
|
|
|
|||||
| Net loss |
$ | (48,912 | ) | $ | (25,361 | ) | ||
|
|
|
|
|
|||||
| Deemed dividend to Series C redeemable convertible preferred stockholders |
(9,675 | ) | — | |||||
|
|
|
|
|
|||||
| Net loss attributable to common stockholders |
$ | (58,587 | ) | $ | (25,361 | ) | ||
|
|
|
|
|
|||||
| Net loss per share attributable to common stockholders, basic and diluted |
$ | (1,997.58 | ) | $ | (864.71 | ) | ||
|
|
|
|
|
|||||
| Weighted-average common shares outstanding, basic and diluted |
29,329 | 29,329 | ||||||
|
|
|
|
|
|||||
See accompanying notes to the condensed financial statements.
F-29
Table of Contents
ELECTRA THERAPEUTICS, INC.
Condensed Statements of Comprehensive Loss
(Unaudited)
(In thousands)
| SIX MONTHS ENDED JUNE 30, |
||||||||
| 2026 | 2025 | |||||||
| Net loss |
$ | (48,912 | ) | $ | (25,361 | ) | ||
| Unrealized gain (loss) on marketable securities |
(34 | ) | (1 | ) | ||||
|
|
|
|
|
|||||
| Total comprehensive loss |
$ | (48,946 | ) | $ | (25,362 | ) | ||
|
|
|
|
|
|||||
See accompanying notes to the condensed financial statements.
F-30
Table of Contents
ELECTRA THERAPEUTICS, INC.
Condensed Statements of Redeemable Convertible Preferred Stock and Stockholders’ Deficit
(Unaudited)
(In thousands, except share amounts)
| SERIES A REDEEMABLE CONVERTIBLE PREFERRED STOCK |
SERIES B REDEEMABLE CONVERTIBLE PREFERRED STOCK |
SERIES C REDEEMABLE CONVERTIBLE PREFERRED STOCK |
COMMON STOCK | ADDITIONAL PAID-IN CAPITAL |
ACCUMULATED DEFICIT |
OTHER COMPREHENSIVE INCOME (LOSS) |
TOTAL STOCKHOLDERS’ DEFICIT |
|||||||||||||||||||||||||||||||||||||||||
| SHARES | AMOUNT | SHARES | AMOUNT | SHARES | AMOUNT | SHARES | AMOUNT | |||||||||||||||||||||||||||||||||||||||||
| Balance as of December 31, 2025 |
15,065,741 | $ | 34,650 | 6,205,494 | $ | 83,836 | 13,214,580 | $ | 133,116 | 29,329 | $ | — | $ | 9,515 | $ | (172,978 | ) | $ | 21 | $ | (163,442 | ) | ||||||||||||||||||||||||||
| Issuance of Series C redeemable convertible preferred stock, net of issuance costs |
— | — | — | — | 4,918,477 | 59,569 | — | — | — | — | — | — | ||||||||||||||||||||||||||||||||||||
| Deemed dividend to Series C redeemable convertible preferred stockholders |
— | — | — | — | — | — | — | — | (9,675 | ) | — | — | (9,675 | ) | ||||||||||||||||||||||||||||||||||
| Stock-based compensation – profits interest units |
— | — | — | — | — | — | — | — | 214 | — | — | 214 | ||||||||||||||||||||||||||||||||||||
| Stock-based compensation expense – options |
— | — | — | — | — | — | — | — | 2,394 | — | — | 2,394 | ||||||||||||||||||||||||||||||||||||
| Other comprehensive income (loss) |
— | — | — | — | — | — | — | — | — | — | (34 | ) | (34 | ) | ||||||||||||||||||||||||||||||||||
| Net loss |
— | — | — | — | — | — | — | — | — | (48,912 | ) | — | (48,912 | ) | ||||||||||||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
| Balance as of June 30, 2026 |
15,065,741 | $ | 34,650 | 6,205,494 | $ | 83,836 | 18,133,057 | $ | 192,685 | 29,329 | $ | — | $ | 2,448 | $ | (221,890 | ) | $ | (13 | ) | $ | (219,455 | ) | |||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
| SERIES A REDEEMABLE CONVERTIBLE PREFERRED STOCK |
SERIES B REDEEMABLE CONVERTIBLE PREFERRED STOCK |
SERIES C REDEEMABLE CONVERTIBLE PREFERRED STOCK |
COMMON STOCK | ADDITIONAL PAID-IN CAPITAL |
ACCUMULATED DEFICIT |
OTHER COMPREHENSIVE INCOME (LOSS) |
TOTAL STOCKHOLDERS’ DEFICIT |
|||||||||||||||||||||||||||||||||||||||||
| SHARES | AMOUNT | SHARES | AMOUNT | SHARES | AMOUNT | SHARES | AMOUNT | |||||||||||||||||||||||||||||||||||||||||
| Balance as of December 31, 2024 |
15,065,741 | $ | 34,650 | 6,205,494 | $ | 83,836 | — | $ | — | 29,329 | $ | — | $ | 7,741 | $ | (111,003 | ) | $ | 1 | $ | (103,261 | ) | ||||||||||||||||||||||||||
| Stock-based compensation – profits interest units |
— | — | — | — | — | — | — | — | 249 | — | — | 249 | ||||||||||||||||||||||||||||||||||||
| Stock-based compensation expense – options |
— | — | — | — | — | — | — | — | 656 | — | — | 656 | ||||||||||||||||||||||||||||||||||||
| Other comprehensive income (loss) |
— | — | — | — | — | — | — | — | — | — | (1 | ) | (1 | ) | ||||||||||||||||||||||||||||||||||
| Net loss |
— | — | — | — | — | — | — | — | — | (25,361 | ) | — | (25,361 | ) | ||||||||||||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
| Balance as of June 30, 2025 |
15,065,741 | $ | 34,650 | 6,205,494 | $ | 83,836 | — | $ | — | 29,329 | $ | — | $ | 8,646 | $ | (136,364 | ) | $ | — | $ | (127,718 | ) | ||||||||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||||||||||||
See accompanying notes to the condensed financial statements.
F-31
Table of Contents
ELECTRA THERAPEUTICS, INC.
Condensed Statements of Cash Flows
(Unaudited)
(In thousands)
| SIX MONTHS ENDED JUNE 30, | ||||||||
| 2026 | 2025 | |||||||
| Cash flows from operating activities |
||||||||
| Net loss |
$ | (48,912 | ) | $ | (25,361 | ) | ||
| Adjustments to reconcile net loss to net cash used in operating activities: |
||||||||
| Stock-based compensation expenses - profits interest units and options |
2,608 | 905 | ||||||
| Depreciation and amortization |
122 | — | ||||||
| Non-cash interest expense – related party |
— | 327 | ||||||
| Non-cash lease expense |
280 | — | ||||||
| Amortization of premiums and discounts of marketable securities, net |
(540 | ) | — | |||||
| Changes in assets and liabilities: |
||||||||
| Other receivables |
(44 | ) | 13 | |||||
| Prepaid expenses and other current assets |
6,376 | (2,490 | ) | |||||
| Operating lease liabilities |
(104 | ) | — | |||||
| Other assets |
(709 | ) | (937 | ) | ||||
| Accounts payable |
74 | 2,854 | ||||||
| Accounts payable – related party |
— | (79 | ) | |||||
| Accrued liabilities |
1,970 | 202 | ||||||
| Accrued liabilities – related party |
— | (122 | ) | |||||
|
|
|
|
|
|||||
| Net cash used in operating activities |
(38,879 | ) | (24,688 | ) | ||||
|
|
|
|
|
|||||
| Cash flows from investing activities |
||||||||
| Proceeds from maturity of marketable securities |
37,420 | 997 | ||||||
| Purchases of marketable securities |
(14,622 | ) | — | |||||
| Purchase of equipment |
(27 | ) | (116 | ) | ||||
|
|
|
|
|
|||||
| Net cash provided by investing activities |
22,771 | 881 | ||||||
|
|
|
|
|
|||||
| Cash flows from financing activities |
||||||||
| Proceeds from the issuance of convertible promissory notes – related party |
— | 40,000 | ||||||
| Proceeds from the issuance of Series C redeemable convertible preferred stock, net of less than $0.1 million issuance costs |
49,894 | — | ||||||
| Payment of deferred offering costs |
(287 | ) | — | |||||
|
|
|
|
|
|||||
| Net cash provided by financing activities |
49,607 | 40,000 | ||||||
|
|
|
|
|
|||||
| Net increase in cash and cash equivalents |
33,499 | 16,193 | ||||||
| Cash and cash equivalents at the beginning of the period |
33,008 | 14,787 | ||||||
|
|
|
|
|
|||||
| Cash and cash equivalents at the end of the period |
$ | 66,507 | $ | 30,980 | ||||
|
|
|
|
|
|||||
| Supplemental disclosures: |
||||||||
| Cash paid for taxes |
$ | 8 | $ | 3 | ||||
|
|
|
|
|
|||||
| Unpaid deferred offering costs included in accrued expenses and other current liabilities |
$ | 1,634 | $ | — | ||||
|
|
|
|
|
|||||
| Deemed dividend to Series C redeemable convertible preferred stockholders |
$ | 9,675 | $ | — | ||||
|
|
|
|
|
|||||
See accompanying notes to the condensed financial statements.
F-32
Table of Contents
1. Organization and Description of Business
Description of Business
Electra Therapeutics, Inc. (“Electra”, or the “Company”) was incorporated in 2018 and is a late clinical-stage biopharmaceutical company focused on pioneering a new class of precision medicines for the treatment of immune-mediated diseases and cancer. The Company’s novel approach targets signal regulatory proteins (“SIRP”), a family of cell surface receptors whose expression is restricted to specific immune cell populations and increases upon activation, to enable selective depletion of disease-driving cells while preserving normal immune function. The Company’s lead product candidate is ipsoprubart, a novel pan-SIRP monoclonal antibody designed to selectively deplete pathological myeloid cells and T cells via binding to SIRPa/ß1/g. The Company is also advancing ELA822, a novel SIRP g-specific monoclonal antibody designed to selectively deplete activated T cells, with potential applications across chronic T cell-mediated immune and inflammatory diseases.
Unaudited Condensed Financial Information
The accompanying unaudited condensed financial statements as of June 30, 2026 and for the six months ended June 30, 2026 and 2025 have been prepared in accordance with U.S. generally accepted accounting principles (“GAAP”) for interim financial information and pursuant to Article 10 of Regulation S-X of the Securities and Exchange Commission (“SEC”). Accordingly, the accompanying unaudited condensed financial statements do not include all of the information and notes required by GAAP for complete financial statements. The unaudited condensed financial statements reflect all adjustments which, in the opinion of management, are necessary for a fair statement of the results for the periods presented. All such adjustments are of a normal and recurring nature. These interim results are not necessarily indicative of results expected for the full fiscal year or any future interim period. The condensed balance sheet as of December 31, 2025 has been derived from the audited financial statements at that date but does not include all of the information and footnotes required by GAAP for complete financial statements. The unaudited condensed financial statements should be read in conjunction with the audited financial statements and notes thereto.
Liquidity and Going Concern
As of June 30, 2026, the Company had total cash, cash equivalents and marketable securities of $97.7 million. The Company has incurred significant net operating losses from operations and had an accumulated deficit of $221.9 million as of June 30, 2026. During the six months ended June 30, 2026, the Company incurred a net loss of $48.9 million. The Company had no debt obligation outstanding as of June 30, 2026. Additionally, the Company anticipates that its net operating losses and negative operating cash flows will continue to increase for the foreseeable future as it continues to expand its research and development programs. The Company expects that its cash, cash equivalents and marketable securities as of June 30, 2026 will not be sufficient to fund its current business plan including related operating expenditure requirements through at least 12 months from the date of issuance of these condensed financial statements. These conditions and events raise substantial doubt about the Company’s ability to continue as a going concern.
The Company’s long-term success is dependent upon its ability to successfully develop and market its pharmaceutical candidates, obtain additional capital when needed and, ultimately, to achieve profitable operations.
The Company plans to raise additional capital to continue funding its development efforts and operations. The Company is actively pursuing raising additional capital; however, there can be no assurance that funds will be available to the Company on acceptable terms, or on a timely basis, if at all based on both market conditions and management’s control. If the Company is unable to obtain an adequate level of capital needed to continue its research and development activities, the Company will need to delay, reduce or eliminate some or all of its planned activities and reduce costs. Doing so will likely have an adverse effect on the ability to execute the Company’s business plan. It is not considered probable that the Company’s plans to raise additional capital nor reduce discretionary spending will alleviate the substantial doubt regarding its ability to continue as a going concern.
The accompanying condensed financial statements do not include any adjustments relating to the recoverability and classification of recorded asset amounts or the amounts and classification of liabilities that might result from the outcome of these uncertainties.
F-33
Table of Contents
Reverse Stock Split
On September 10, 2026, the Company filed a certificate of amendment to the amended and restated certificate of incorporation to effect a 1-for-1.5002 reverse stock split of the Company’s outstanding shares of common stock and redeemable preferred stock. Upon the effectiveness of the reverse stock split, (i) all shares of outstanding common stock were adjusted; (ii) all shares of outstanding redeemable convertible preferred stock were adjusted; (iii) the number of shares of common stock for each outstanding option to purchase common stock was adjusted; and (iv) the exercise price of each outstanding option to purchase common stock was adjusted. All of the outstanding common stock share numbers (including shares of common stock subject to the Company’s options and as converted for the outstanding convertible preferred stock), share prices, exercise prices and per share amounts contained in the financial statements have been retroactively adjusted in the financial statements to reflect this reverse stock split for all periods presented. The par value per share and the authorized number of shares of common stock and preferred stock were not adjusted as a result of the reverse stock split.
Risks and Uncertainties
The Company is subject to risks and uncertainties common to early-stage companies in the biopharmaceutical industry, including, but not limited to, successful discovery and development of its product candidates, development by competitors of new technological innovations, dependence on key personnel, the ability to attract and retain qualified employees, protection of proprietary technology, compliance with governmental regulations, the change of internal trade relations, new legislation and tariffs, and the ability to secure additional capital to fund operations and commercial success of its product candidates. Ipsoprubart and any future product candidates the Company may develop will require extensive nonclinical and clinical testing and regulatory approval prior to commercialization. These efforts require significant amounts of additional capital, adequate personnel, and infrastructure and extensive compliance-reporting capabilities. Even if the Company’s product development efforts are successful, it is uncertain when, if ever, the Company will realize significant revenue from product sales.
2. Summary of Significant Accounting Policies
The Company’s significant accounting policies are disclosed in the audited financial statements for the years ended December 31, 2025 and 2024, included elsewhere in this prospectus. Since the date of those annual financial statements, there have been no changes to the Company’s significant accounting policies, except as noted below.
Deferred Offering Costs
The Company capitalizes certain legal, professional accounting and other third-party fees that are directly associated with the in-process equity financing as deferred offering costs until such financing is consummated. After consummation of such equity financing, these costs are recorded as a reduction of the proceeds generated as a result of the offering within additional paid in capital. In the event the planned equity financing is abandoned, the deferred offering costs will be reclassified to general and administrative expenses in the Company’s statements of operations. The Company did not record any deferred issuance costs as of December 31, 2025, and recorded deferred offering costs of $1.9 million within other assets, non-current as of June 30, 2026.
Performance-Based Options
The Company has issued stock options that vest in conjunction with performance conditions. For performance-based options, the Company recognizes share-based compensation expense over the requisite service period when achievement of the performance criteria is considered probable based on the Company’s best estimate at the end of each reporting period.
3. Related Party Transactions
Intercompany Services Agreements with Star Therapeutics, Inc.
Prior to January 1, 2024, the Company did not have any employees and the Company’s operations were conducted by employees of a subsidiary of Star Therapeutics LLC (“Star LLC”), Star Therapeutics, Inc. (“Star Inc.”), Electra’s parent company through June 2023. These related party transactions were governed by the Mutual Intercompany Services Agreements (“ISA”) between the Company and Star Inc., through its termination on October 31, 2025. Per the ISA, the Company bore all expenses incurred by Star Inc. in performing its duties in accordance with the agreements and in managing and operating the businesses of Electra with a minimal mark-up on expenses. Accordingly, expenses incurred and paid through the affiliated entity’s operating account on behalf of Electra were
F-34
Table of Contents
reimbursed by the Company to Star Inc. Operating expenses incurred by the affiliated entity included but were not limited to: service provider employee compensation expenses and other related overhead such as rent and general office expenses, utilities, information technology services, insurance, human resource management, taxes and other personnel costs.
The Company incurred $0.7 million of expenses under the ISA during the six months ended June 30, 2025, which were reimbursed by the Company to Star Inc. The Company did not incur expenses under the ISA during the six months ended June 30, 2026 due to its termination on October 31, 2025. As of June 30, 2026 and December 31, 2025, there were no amounts due to or from Star Inc. related to the ISA.
Convertible Promissory Notes
On March 7, 2025, the Company issued convertible promissory notes with an aggregate principal amount of $20.0 million (the “Initial Notes”) to existing investors of the Company’s redeemable convertible preferred stock. On June 30, 2025, the Company issued additional convertible promissory notes with an aggregate principal amount of $20.0 million (the “Subsequent Notes” and, together with the Initial Notes, the “Notes”) to existing investors of the Company’s redeemable convertible preferred stock. The Notes bore interest at 8% per annum and were scheduled to mature on June 30, 2026. However, on October 15, 2025, the Company completed a qualified preferred stock equity financing through the issuance of Series C redeemable convertible preferred stock (the “Series C Preferred Stock”). In accordance with their original terms, the outstanding principal of the Notes and all accrued and unpaid interest, totaling $41.4 million, automatically converted into 4,080,265 shares of Series C Preferred Stock at a conversion price of $10.15 per share on that date. The conversion price equaled the cash price per share paid by unrelated third-party investors participating in the Series C Preferred Stock financing. As of June 30, 2026 and December 31, 2025, no convertible promissory notes remained outstanding (see Note 8 – Convertible Notes – Related Party and Note 10 – Redeemable Convertible Preferred Stock).
4. Fair Value Measurements
On a recurring basis, the Company measures certain financial assets and liabilities at fair value, including the Company’s cash equivalents and marketable securities. The following tables summarize the Company’s assets measured at fair value on a recurring basis, by level, within the fair value hierarchy (in thousands):
| FAIR VALUE AS OF JUNE 30, 2026 | ||||||||||||||||
| LEVEL 1 | LEVEL 2 | LEVEL 3 | TOTAL | |||||||||||||
| Cash equivalents |
||||||||||||||||
| Money market funds |
$ | 66,011 | $ | — | $ | — | $ | 66,011 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total cash equivalents |
66,011 | — | — | 66,011 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Marketable securities |
||||||||||||||||
| U.S. Government securities |
— | 8,508 | — | 8,508 | ||||||||||||
| Agency discount notes |
— | 1,991 | — | 1,991 | ||||||||||||
| Commercial paper |
— | 12,653 | — | 12,653 | ||||||||||||
| Corporate securities |
— | 5,558 | — | 5,558 | ||||||||||||
| Asset backed securities |
— | 2,508 | — | 2,508 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total marketable securities |
— | 31,218 | — | 31,218 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total assets measured at fair value |
$ | 66,011 | $ | 31,218 | $ | — | $ | 97,229 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
F-35
Table of Contents
| FAIR VALUE AS OF DECEMBER 31, 2025 | ||||||||||||||||
| LEVEL 1 | LEVEL 2 | LEVEL 3 | TOTAL | |||||||||||||
| Cash equivalents |
||||||||||||||||
| Money market funds |
$ | 26,517 | $ | — | $ | — | $ | 26,517 | ||||||||
| Agency discount notes |
— | 1,998 | — | 1,998 | ||||||||||||
| Commercial paper |
— | 3,995 | — | 3,995 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total cash equivalents |
26,517 | 5,993 | — | 32,510 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Marketable securities |
||||||||||||||||
| U.S. Government securities |
— | 8,988 | — | 8,988 | ||||||||||||
| Agency discount notes |
— | 5,959 | — | 5,959 | ||||||||||||
| Commercial paper |
— | 29,559 | — | 29,559 | ||||||||||||
| Corporate securities |
— | 9,004 | — | 9,004 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total marketable securities |
— | 53,510 | — | 53,510 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total assets measured at fair value |
$ | 26,517 | $ | 59,503 | $ | — | $ | 86,020 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
Money market funds are included within Level 1 of the fair value hierarchy because they are valued using quoted market prices. Agency discount notes, asset-backed securities, commercial paper, corporate securities and U.S. government securities are classified within Level 2 of the fair value hierarchy as they take into consideration valuations obtained from third-party pricing services. The pricing services utilize industry standard valuation models, including both income-based and market-based approaches, for which all significant inputs are observable, either directly or indirectly, to estimate the fair value. These inputs include reported trades of and broker/dealer quotes on similar securities, issuer credit spreads, benchmark securities, prepayment or default projections based on historical data and other observable inputs. The Company did not have any Level 3 securities, and there were no transfers between Level 2 or 3, during the six months ended June 30, 2026 or for the year ended December 31, 2025. All debt securities are Level 2 and classified as available-for-sale.
As of June 30, 2026 and December 31, 2025, the Company had no financial liabilities that required fair value measurement. The carrying values of other current assets, accounts payable and accrued expenses approximate their fair values due to the short-term nature of these assets and liabilities.
5. Marketable Securities
The following tables summarize the amortized cost and the unrealized gains (losses) of the marketable securities presented within cash equivalents and marketable securities (in thousands):
| JUNE 30, 2026 | ||||||||||||||||
| AMORTIZED COST |
UNREALIZED GAIN |
UNREALIZED (LOSS) |
ESTIMATED FAIR VALUE |
|||||||||||||
| Assets |
||||||||||||||||
| Agency discount notes |
$ | 1,993 | $ | — | $ | (2 | ) | $ | 1,991 | |||||||
| Commercial paper |
12,656 | — | (3 | ) | 12,653 | |||||||||||
| Corporate securities |
5,560 | — | (2 | ) | 5,558 | |||||||||||
| Asset-backed securities |
2,510 | — | (2 | ) | 2,508 | |||||||||||
| U.S. Government securities |
8,512 | — | (4 | ) | 8,508 | |||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total assets measured at fair value |
$ | 31,231 | $ | — | $ | (13 | ) | $ | 31,218 | |||||||
|
|
|
|
|
|
|
|
|
|||||||||
F-36
Table of Contents
| DECEMBER 31, 2025 | ||||||||||||||||
| AMORTIZED COST |
UNREALIZED GAIN |
UNREALIZED (LOSS) |
ESTIMATED FAIR VALUE |
|||||||||||||
| Assets |
||||||||||||||||
| Agency discount notes |
$ | 7,955 | $ | 2 | $ | — | $ | 7,957 | ||||||||
| Commercial paper |
33,546 | 8 | — | 33,554 | ||||||||||||
| Corporate securities |
9,000 | 4 | — | 9,004 | ||||||||||||
| U.S. Government securities |
8,982 | 6 | — | 8,988 | ||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total assets measured at fair value |
$ | 59,483 | $ | 20 | $ | — | $ | 59,503 | ||||||||
|
|
|
|
|
|
|
|
|
|||||||||
As of June 30, 2026, the weighted-average remaining maturity of the Company’s marketable securities was less than one year. The Company believes that an allowance for credit losses is unnecessary. As of June 30, 2026, the Company held 17 marketable securities in an unrealized loss position. All investments with unrealized losses have been in a loss position for less than twelve months, and the unrealized losses were considered to be temporary in nature. The Company does not intend to sell the investments in an unrealized loss position, and it is not more likely than not that the Company will be required to sell the investments before recovery of their amortized cost basis. To date, the Company has not recorded any impairment charges on available-for-sale securities. Additionally, accretion income due and accrued as of June 30, 2026 and December 31, 2025 of $0.5 million and $0.1 million, respectively, are recorded as components of the marketable securities on the Company’s condensed balance sheets.
6. Balance Sheet Components
Prepaid Expenses and Other Current Assets
Prepaid expenses and other current assets consisted of the following (in thousands):
| JUNE 30, 2026 |
DECEMBER 31, 2025 |
|||||||
| Prepaid manufacturing |
$ | 798 | $ | 8,516 | ||||
| Prepaid clinical |
1,112 | — | ||||||
| Prepaid insurance |
70 | 53 | ||||||
| Prepaid deposits and other |
392 | 179 | ||||||
|
|
|
|
|
|||||
| Total prepaid expenses |
$ | 2,372 | $ | 8,748 | ||||
|
|
|
|
|
|||||
Accrued Liabilities
Accrued liabilities consisted of the following (in thousands):
| JUNE 30, 2026 |
DECEMBER 31, 2025 |
|||||||
| Accrued clinical trial costs |
$ | 2,328 | $ | 1,706 | ||||
| Accrued manufacturing |
1,952 | 566 | ||||||
| Accrued research and development |
622 | 429 | ||||||
| Accrued bonus |
1,408 | 2,014 | ||||||
| Accrued general and administrative |
1,883 | 330 | ||||||
|
|
|
|
|
|||||
| Total accrued liabilities |
$ | 8,193 | $ | 5,045 | ||||
|
|
|
|
|
|||||
F-37
Table of Contents
7. Leases
Effective September 10, 2025, the Company entered into a noncancelable operating lease agreement for approximately 14,055 rentable square feet of office and laboratory space located in South San Francisco, California (the “Premises”). The Lease has a three-year term expiring on September 30, 2028, and includes an initial rent abatement period. The Lease provides the Company with two consecutive options to extend the lease term for three years each, at the then-prevailing fair rental value; the Company has not included these extension options in the lease term as it is not reasonably certain to exercise either option. In addition to base rent, the Company is required to pay its proportionate share of the building’s operating and tax expenses, which are accounted for as variable lease payments and are not included in the measurement of the lease liability. The Company used an incremental borrowing rate of 6.45% to discount the lease payments, as the rate implicit in the Lease was not readily determinable. No lease expense was recorded for the six months ended June 30, 2025.
The components of lease expense were as follows (in thousands):
| SIX MONTHS ENDED JUNE 30, 2026 |
||||
| Operating lease cost (straight-line) |
$ | 422 | ||
| Variable lease cost (direct expenses) |
223 | |||
|
|
|
|||
| Total lease cost |
$ | 645 | ||
|
|
|
|||
Operating lease cost is recognized on a straight-line basis over the lease term. Cash paid for amounts included in the measurement of lease liabilities for the six months ended June 30, 2026 and 2025 were $0.2 million and zero, respectively, and were included in net cash used in operating activities in the Company’s statements of cash flows.
Maturities of operating lease liabilities as of June 30, 2026 were as follows (in thousands):
| AMOUNT | ||||
| 2026 (remaining) |
$ | 459 | ||
| 2027 |
942 | |||
| 2028 |
725 | |||
|
|
|
|||
| Total operating lease payments |
2,126 | |||
| Less: imputed interest |
(143 | ) | ||
|
|
|
|||
| Total operating lease liabilities |
$ | 1,983 | ||
|
|
|
|||
8. Convertible Notes – Related Party
On March 7, 2025, the Company issued Initial Notes to multiple investors. The Initial Notes bore interest at an annual rate of 8% per annum on the outstanding principal balance. Interest commenced on the closing date of March 7, 2025 and was computed on the basis of a year of 365 days for the actual number of days elapsed. The outstanding principal amount and all accrued and unpaid interest was due and payable upon demand by the Holders on or after June 30, 2026 (the “Maturity Date”). Each Holder committed to purchase additional notes in one or more future closings with the combined aggregate commitment of all Holders not to exceed $20.0 million.
On June 30, 2025, the Company issued the Subsequent Notes in the aggregate principal amount of $20.0 million pursuant to the commitment. The Subsequent Notes had terms identical to the Initial Notes, except for their respective principal amounts. The aggregate principal amount of the Notes outstanding was $40.0 million. The Notes were issued to existing investors of the Company’s redeemable convertible preferred stock, therefore, the issuances are considered related party transactions.
The Notes contained the following settlement provisions: (1) Upon the occurrence of a qualified preferred stock equity financing (a “Qualified Financing”) on or before the Maturity Date, the amount due automatically converts
F-38
Table of Contents
into fully paid and nonassessable shares of the preferred stock issued in such Qualified Financing at a price per share equal to the cash price per share paid by the other purchasers in the Qualified Financing; (2) if the Company completed a preferred stock equity financing that raised less than the Qualified Financing threshold of proceeds totaling $40.0 million, excluding note conversions, the Holders holding a majority of the principal outstanding under the Notes could elect to treat such financing as a Qualified Financing, with the amount due converting into preferred stock on the same terms, or hold the Notes until maturity; (3) upon the occurrence of a change of control, Holders were entitled to receive a cash payment equal to the amount due plus an applicable premium (the applicable premium was 20% if the change of control occurred prior to March 7, 2026, 50% if on or after March 7, 2026, and 100% premium if on or after the Maturity Date); and (4) upon an event of default, the amount due would become immediately due and payable in cash.
The Company elected the fair value option under ASC 825-10 to account for the Notes at the time of their initial recognition. The convertible notes are remeasured at fair value at the end of each reporting period, until fully repaid or converted, using the discounted cash flow method and include unobservable inputs derived from management’s estimates and assumptions. Management’s estimates and assumptions are based in part on external data and internal data and involve a significant degree of judgement. The primary unobservable inputs, classified as Level 3 under the fair value hierarchy, are the probability and timing of Qualified Financing and change of control events. The discount rate utilized in the fair value model was approximately 21%.
Transaction costs incurred in connection with the issuance of the Notes were expensed as incurred. Interest expense from the Notes is presented in the statement of operations and comprehensive loss. For the year ended December 31, 2025, the Company recognized $1.4 million of expense in the statement of operations and comprehensive loss for the difference between the proceeds received and the fair value of the Notes, which was due to accrued interest on the Notes as of the conversion date. On issuance, total issuance costs of $0.1 million were expensed and recognized as general and administrative expense in the statement of operations and comprehensive loss.
On October 15, 2025, the Company completed a Qualified Financing through the issuance of Series C Preferred Stock (see Note 10 – Stockholders’ Deficit). Upon the closing of the Qualified Financing, the outstanding principal amount of the Notes and all accrued and unpaid interest thereon, totaling $41.4 million, were automatically converted into 4,080,265 shares of Series C Preferred Stock at a conversion price of $10.15 per share.
As of June 30, 2026 and December 31, 2025, no convertible promissory notes remained outstanding. There was no gain or loss on extinguishment in the statement of operations for the periods ended June 30, 2026 or December 31, 2025. For the six months ended June 30, 2025, the Company recognized $0.3 million of interest expense in the statement of operations for the difference between the proceeds received and the fair value of the Notes. No interest expense was recorded for the six months ended June 30, 2026.
9. Commitments and Contingencies
Commitments
The Company enters into contractual agreements with various suppliers in the normal course of its business. All contracts are terminable, with varying provisions regarding termination. If a contract with a specific vendor were to be terminated, the Company would only be obligated for the products or services that the Company had received through the time of termination.
In September 2025, the Company entered into a noncancelable operating lease agreement for approximately 14,055 rentable square feet of office and laboratory space in South San Francisco, California (see Note 7 – Leases). The lease provides for escalating annualized base rent payments starting at $0.9 million and increasing to $1.0 million in the final year of the lease. Remaining lease payments from July 1, 2026 through the end of the lease term total $2.1 million.
The Company is also party to a mouse platform agreement with an unrelated third party. Under the agreement, the Company was granted non-exclusive licenses to use transgenic mice to generate antibodies and to exploit products containing the resulting antibodies (“Antibody Products”). Under the agreement, the Company paid a one-time
F-39
Table of Contents
initial technology access fee and is obligated to pay annual low five-digit maintenance fees per distinct antigen with which the Company immunized the mice, continuing until the first commercial sale of an Antibody Product. In addition, the agreement requires the Company to make certain development and commercial milestone payments upon the achievement of specified events by each Antibody Product, as well as pay royalties on net sales of Antibody Products. Aggregate milestone payments are up to the low double-digit millions of dollars per distinct Antibody Product. Royalties under the agreement are a low single digit percentage of net sales of all Antibody Products and are payable on a product-by-product and country-by-country basis beginning with the first commercial sale of the applicable Antibody Product in the applicable country and continuing for 10 years thereafter. The agreement remains in effect until the expiration of the last-to-expire royalty term, unless earlier terminated by the Company at will or by either party for the other party’s uncured material breach of a material provision of the agreement.
Contingencies
From time to time, the Company may have certain contingent liabilities that arise in the ordinary course of its business activities. The Company accrues a liability for such matters when it is probable that future expenditures will be made and that such expenditures can be reasonably estimated. Significant judgment is required to determine both probability and the estimated amount. There are no matters pending that the Company currently believes are reasonably possible or probable of having a material impact to the Company’s financial position, results of operations, or statements of cash flows.
In the normal course of operations, the Company may become involved in various legal proceedings. As of June 30, 2026 and December 31, 2025, the Company had not recorded accruals for probable losses related to any existing or pending litigation as the Company’s management has determined that there are no matters where a potential loss is probable and reasonably estimable. The Company does not believe that any existing or pending claims would have a material impact on the Company’s financial position, results of operations, or statements of cash flows.
Indemnification
In the normal course of business, the Company enters into contracts and agreements that contain a variety of representations and warranties and may provide for indemnification of the counterparty. The Company’s exposure under these agreements is unknown because it involves claims that may be made against it in the future but have not yet been made.
In accordance with the Company’s Amended and Restated Certificate of Incorporation (the “Certificate of Incorporation”) and Amended and Restated Bylaws, the Company has indemnification obligations to its officers and directors, subject to some limits, with respect to their service in such capacities. The Company has also entered into indemnification agreements with its directors and certain of its officers. To date, the Company has not been subject to any claims, and it maintains director and officer insurance that may enable it to recover a portion of any amounts paid for future potential claims.
The Company’s exposure under these agreements is unknown because it involves claims that may be made against it in the future but have not yet been made. The Company believes that the fair value of these indemnification obligations is minimal; accordingly, it has not recognized any liabilities relating to these obligations for any period presented.
10. Redeemable Convertible Preferred Stock
Although the Company’s redeemable convertible preferred stock is not mandatorily or currently redeemable, a liquidation or winding up of the Company could constitute an event outside its control. Therefore, all shares of redeemable convertible preferred stock have been presented outside of permanent equity due to being contingently redeemable. The redeemable convertible preferred stock is redeemable for cash or other assets upon the occurrence of a deemed liquidation event (including a change of control), which constitutes an event not solely within the Company’s control. Accordingly, the redeemable convertible preferred stock is classified outside of permanent equity and is presented as temporary equity in the balance sheets.
F-40
Table of Contents
The following table presents information related to redeemable convertible preferred stock (in thousands, except share and per share amounts):
| AS OF JUNE 30, 2026 | ||||||||||||||||||||
| SHARES AUTHORIZED |
SHARE ISSUED AND OUTSTANDING |
LIQUIDATION PREFERENCE |
CARRYING AMOUNT |
ORIGINAL ISSUE PRICE |
||||||||||||||||
| Series A |
22,601,626 | 15,065,741 | $ | 34,750 | $ | 34,650 | $ | 2.31 | ||||||||||||
| Series B |
9,309,492 | 6,205,494 | 84,019 | 83,836 | $ | 13.54 | ||||||||||||||
| Series C |
27,203,243 | 18,133,057 | 184,133 | 192,685 | $ | 10.15 | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||||||
| Total redeemable convertible preferred stock |
59,114,361 | 39,404,292 | $ | 302,902 | $ | 311,171 | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||||||
On October 15, 2025, pursuant to a Series C Preferred Stock Purchase Agreement (the “Purchase Agreement”), the Company issued 9,134,315 shares of its Series C Preferred Stock at a purchase price of $10.15 per share (the “Original Issue Price”) to multiple investors (the “Investors”) in a transaction referred to as the “Initial Closing”, resulting in gross proceeds of $92.8 million.
At the Initial Closing, previously issued Notes with aggregate principal and accrued interest of $41.4 million were automatically converted into 4,080,265 shares of Series C Preferred Stock at a conversion price equal to the Original Issue Price. $69.2 million of the $134.2 million Series C Preferred Stock was purchased by investors who had invested in the Company prior to the Initial Closing, and are considered to be related parties.
On June 18, 2026, pursuant to the Series C Preferred Stock Purchase Agreement, the Company closed on a subsequent closing (the “Second Tranche Closing”) and issued shares of its Series C Preferred Stock at the Original Issue Price of $10.15 per share on the same terms as the Initial Closing. The Second Tranche Closing was made up of the same investors as those who participated in the Initial Closing in October 2025 and were committed to the Second Tranche. The Second Tranche Closing resulted in the issuance of 4,918,477 shares of Series C Preferred Stock for total gross proceeds of $49.9 million. $14.9 million of the $49.9 million Series C Preferred Stock was purchased by investors who had invested in the Company prior to the Initial Closing, and are considered to be related parties. Due to the terms and price being the same as the Initial Closing and the passage of time between the two closing, a deemed dividend was recorded to additional paid-in capital for $9.7 million upon the issuance of the shares of Series C Preferred Stock at the Second Tranche Closing which represents the aggregate fair value of the shares of Series C Preferred Stock issued in excess of cash proceeds received.
The Series C Preferred Stock and the associated rights for the Second Tranche Closing were contractually linked and unable to be sold or exercised separately except in limited affiliate transfer scenarios. As such, the Second Tranche Closing was defined as an embedded feature within the Series C Preferred Stock financing rather than a freestanding instrument, and no liability was recorded for the Second Tranche Closing in 2025.
Dividend: Each holder of redeemable convertible preferred stock is entitled to receive non-cumulative dividends at the rate of 8% per annum for each share of redeemable convertible preferred stock outstanding, when, as and if declared by the Company’s Board of Directors (the “Board of Directors”). These dividends are payable in preference to common stock dividends. To date, the Company has not declared or paid any dividends.
Liquidation: In the event of any liquidation event, either voluntary or involuntary, the holders of preferred stock shall be entitled to receive out of the proceeds or assets available for distribution, prior and in preference to any distributions to the holders of common stock, an amount per share equal to the sum of the applicable original issuance price, plus declared but unpaid dividends on such share. If the proceeds distributed among the holders of the preferred stock are insufficient to permit the payment to such holders, then the entire proceeds available for distribution shall be distributed ratably among the holders of preferred stock in the same proportion to respective amounts, which would be payable if paid in full.
Conversion for Series A and B Preferred Stock: Each share of preferred stock shall be convertible, at the option of the holder, at any time after the date of issuance of such share into such number of shares of common stock
F-41
Table of Contents
determined by dividing the original issue price for preferred stock series by the conversion price. The conversion price is the same as the original issuance price for Series A Preferred Stock and Series B Preferred Stock.
Conversion of each series of redeemable convertible preferred stock into common stock is automatic upon the earlier of: (1) the closing of the sale of common stock in a firm commitment underwritten public offering that results in at least $100 million of gross proceeds at a minimum price per share of $10.15, subject to adjustment in the event of a stock split; or (2) the date, or the occurrence of an event, specified by vote or written consent or agreement of the Requisite Investors (“Requisite Investors” means Investors holding at least 70% of the shares of Series B Preferred Stock and Series C Preferred Stock then outstanding (voting together as a single class and not as separate series, and on an as-converted basis).
Conversion for Series C Preferred Stock: Each share of preferred stock shall be convertible, at the option of the holder, at anytime after the earlier of: (1) the day after the Second Tranche Closing; (2) a Liquidation Event; (3) the termination of the Second Tranche obligation; or (4) another date designated by the Company and the Requisite Investors, excluding any shares held by defaulting investors.
The conversion price is the same as the original issuance price for Series C Preferred Stock.
Conversion of each series of redeemable convertible preferred stock into common stock is automatic upon the earlier of: (1) the closing of the sale of common stock in a firm commitment underwritten public offering that results in at least $100.0 million of gross proceeds at a minimum price per share of $10.15, subject to adjustment in the event of a stock split; or (2) the date, or the occurrence of an event, specified by vote or written consent or agreement of the Requisite Investors.
The conversion price is also subject to standard anti-dilutive adjustments for stock splits, combinations, dividends, distributions, reclassifications, and recapitalizations, as set forth in the Certificate of Incorporation. Any downward adjustment to the conversion price of the Series C Preferred Stock may be waived, prospectively or retroactively, by the consent or vote of holders of at least 60% of the then-outstanding shares of Series C Preferred Stock (voting as a single class on an as-converted basis).
Voting Rights: The holder of each share of preferred stock shall have the right to one vote for each share of common stock into which shares of preferred stock could then be converted. The Board of Directors of the Company is set at 7 members with 1 vacancy, appointed as follows: 2 directors from the Holders of Series B Preferred Stock, 1 director from the Holders of Series C Preferred Stock, 4 directors from the Holders of a majority of outstanding capital stock (voting together as a single class on an as-converted basis).
As of June 30, 2026 and December 31, 2025, a deemed liquidation event was not considered probable of occurring. Accordingly, the redeemable convertible preferred stock has not been remeasured to its redemption value. The Company will reassess the probability of a deemed liquidation event and the classification and measurement of the redeemable convertible preferred stock at each reporting period end.
11. Stockholders’ Deficit
Stock-Based Compensation
The Company recognized stock-based compensation as follows (in thousands):
| SIX MONTHS ENDED JUNE 30, | ||||||||
| 2026 | 2025 | |||||||
| Research and development expense – PI Units |
$ | 50 | $ | 66 | ||||
| Research and development expense – options |
1,157 | 371 | ||||||
|
|
|
|
|
|||||
| Total research and development expense |
1,207 | 437 | ||||||
| General and administrative expense – PI Units |
164 | 183 | ||||||
| General and administrative expense – options |
1,237 | 285 | ||||||
|
|
|
|
|
|||||
| Total general and administrative expense |
1,401 | 468 | ||||||
|
|
|
|
|
|||||
| Total stock-based compensation expense |
$ | 2,608 | $ | 905 | ||||
|
|
|
|
|
|||||
F-42
Table of Contents
Profits Interest Units
In December 2018, Star LLC adopted the 2018 Equity Incentive Plan (the “Star LLC Plan”) pursuant to which it could grant profits interest unit (“PI Units”) awards to its employees, managers, and consultants, including those hired and engaged by its affiliated entities. In June 2023, Star LLC’s members formed Electra Therapeutics LLC (“Electra LLC”) and contributed their Series A redeemable convertible preferred shares to Electra LLC in exchange for the Series B preferred units, Series A preferred units, Series Seed preferred units, and common units of Electra LLC. This transaction transferred all of Star LLC’s equity ownership in the Company to Electra LLC, a sister company of Star LLC immediately after the spin-off. In connection with the spin-off, Electra LLC adopted the 2023 Equity Incentive Plan (the “Electra LLC Plan”) and granted PI Units to Star LLC’s PI Unit holders who supported the operations of Electra through the ISA. On December 31, 2023, Star LLC terminated 16 employees who were immediately employed by Electra, effective January 1, 2024. On January 1, 2024, the 4.5 million PI Units previously awarded to these holders for services rendered under the MISA were remeasured with a weighted-average fair value of $1.04 per PI Unit. All other terms of the PI Units remained the same.
Under the provisions of the Electra LLC Plan, the board of managers of Electra LLC (the “Board of Managers”) may grant PI Units to its employees, managers, and consultants, including those hired and engaged by its affiliated entities (collectively, the “Participants”). PI Units are common units issued to Participants with a threshold amount. In the event of distribution by Electra LLC, the proceeds distributed to the holder would be reduced by the threshold amount. PI Units are economically similar to a stock option award and vest based on time or performance-based milestones, as determined by the Board of Managers of Electra LLC and stipulated in the grant agreements.
PI Units generally vest 25% after one year, with the remainder vesting monthly over the following three-year period. The underlying terms and intended purpose of the PI Units are more akin to stock-based compensation for employees and non-employees than a performance bonus or profit-sharing arrangement.
No PI Units were granted during the six months ended June 30, 2026 and the year ended December 31, 2025.
Stock Incentive Plan
In 2022, the Board of Directors approved the adoption of the 2022 Stock Plan (the “Plan”). The Plan allows for the award of incentive and non-qualified stock options and restricted stock to employees, outside directors, and consultants of the Company. In February 2026, the Board of Directors approved an amendment of the Plan to increase the number of shares of Common Stock reserved for issuance under the Plan from 5,663,057 shares to 6,196,109 shares. As of June 30, 2026, 464,837 shares were available for award under the Plan. The Board of Directors administers the Plan and determines the exercise price of options, purchase price for restricted stock, the rates at which awards vest, and the other terms and conditions of the awards. Options and restricted stock generally vest 25% upon the first anniversary of the grant date and an additional 1/48th of the shares vest each month of continuous service thereafter for 36 months, and expire 10 years from the date of grant. Stock options are eligible for early exercise. No tax benefits were realized from options and other share-based payment arrangements during the year.
The Company grants stock options at exercise prices deemed by the Board of Directors to be equal to the fair value of the common stock at the time of grant. The fair value of common stock has been determined by the Board of Directors at each stock option measurement date based on a variety of different factors, including contemporaneous independent valuations performed by an independent third-party valuation firm; the prices per share of redeemable convertible preferred stock sold to investors in arm’s length transactions, and the rights, preferences, and privileges of the redeemable convertible preferred stock relative to the Company’s common stock; the Company’s stage of development and material risks related to the business; results of operations and financial position, including levels of available capital resources; progress of research and development activities; the lack of marketability of the Company’s common stock as a private company; the status of strategic transactions; the hiring of key personnel and the experience of management; the likelihood and timing of achieving a liquidity event, such as an initial public offering or a sale of the Company, given prevailing market conditions; market performance of comparable publicly traded companies; trends and developments in the life sciences and biopharmaceutical industry; and external market conditions affecting the industry sector.
F-43
Table of Contents
The Company calculated the value of the stock options using the Black-Scholes option pricing model with the following assumptions:
| SIX MONTHS ENDED JUNE 30, | ||||||||
| 2026 | 2025 | |||||||
| Risk-free interest rate |
3.6%–4.2% | 4.4%–4.4% | ||||||
| Expected term, in years |
5.00–6.18 | 6.08 | ||||||
| Expected dividend yield |
— | — | ||||||
| Expected volatility |
103.3%–131.8% | 93.1%–93.1% | ||||||
The Black-Scholes option pricing model requires the use of subjective assumptions to determine the fair value of stock-based awards, including:
| ∎ | Risk-Free Interest Rate – based on the yield available on U.S. Treasury zero-coupon issues similar in duration to the expected term of the award. |
| ∎ | Expected Term – using the simplified method which calculates the expected term as the average of the time-to-vesting and the contractual life of the stock options. |
| ∎ | Expected Dividend Yield – assuming the expected dividend yield is zero, as the Company has no history or expectation of paying cash dividends on common stock in the foreseeable future. |
| ∎ | Expected Volatility – based on a study of publicly traded industry peer companies which were identified based on industry, stage of development, size, and financial leverage of potential comparable companies. Historical volatility is calculated based on a period of time commensurate with the expected term assumption. |
The Company records stock-based compensation expense on options as they vest and records forfeitures as they occur.
Performance-based Options
On February 13, 2026, the Company granted 364,802 performance-based options to certain officers. The award has an exercise price of $4.96 per share and vests ratably on a monthly basis over four years, subject to and contingent on the achievement of specified operational milestones approved by the Board of Directors. As of June 30, 2026, the requisite performance criteria for these options was not achieved and not probable. Therefore, no compensation expense was recognized during the six months ended June 30, 2026.
The following table summarizes the Company’s stock option activity, including performance-based options:
| OPTIONS OUTSTANDING | ||||||||||||
| NUMBER OF OPTIONS |
WEIGHTED- AVERAGE EXERCISE PRICE PER SHARE |
WEIGHTED- AVERAGE REMAINING CONTRACTUAL TERM (IN YEARS) |
||||||||||
| Balances as of December 31, 2025 |
2,266,067 | $ | 4.86 | 8.05 | ||||||||
|
|
|
|
|
|
|
|||||||
| Options granted |
3,490,119 | $ | 4.96 | |||||||||
| Options forfeited |
(54,243 | ) | $ | 4.96 | ||||||||
| Options cancelled |
— | $ | — | |||||||||
|
|
|
|||||||||||
| Balances as of June 30, 2026 |
5,701,943 | $ | 4.92 | 8.73 | ||||||||
|
|
|
|||||||||||
| Vested as of June 30, 2026 |
1,580,536 | |||||||||||
| Expected to vest as of June 30, 2026 |
4,121,407 | |||||||||||
F-44
Table of Contents
As of June 30, 2026, the total gross unrecognized stock-based compensation expense related to unvested stock options aggregated to $15.9 million. These expenses are expected to be recognized over a weighted-average period of 3.0 years. The total fair value of options vested during the six months ended June 30, 2026 was $1.9 million.
12. Segments
The Company’s CODM is its Chief Executive Officer, who reviews financial information presented on a net loss basis as reported in the statement of operations in order to make decisions about allocating resources and assessing performance for the entire Company. The CODM also utilizes the Company’s annual budget and strategic plan, as a key input to resource allocation. The CODM function approves of key operating and strategic decisions. The CODM function views the Company’s operations and manages its business as a single reportable operating segment.
The Company monitors its cash, cash equivalents and investments as reported on the Company’s balance sheets to determine funding needed for furthering its research and development activities, while continuing general and administrative operations. The CODM function is regularly provided with the following significant segment expenses. Significant expenses include research and development and general and administrative expenses, which are each separately presented in the Company’s statements of operations. The CODM reviews significant expenses within both the research and development and the general and administrative categories. Other segment items within net loss include interest income, interest expense, and other income (expense). No product revenue has been generated since inception and all assets are held in the United States. See the financial statements for other financial information regarding the Company’s operating segment. The CODM is regularly provided with the following significant segment expenses:
| SIX MONTHS ENDED JUNE 30, | ||||||||
| 2026 | 2025 | |||||||
| Operating expenses: |
||||||||
| Research and development: |
||||||||
| Direct program costs: |
||||||||
| Ipsoprubart |
$ | 26,124 | $ | 8,154 | ||||
| Other programs |
7,546 | 5,385 | ||||||
| Unallocated costs: |
||||||||
| Personnel-related |
6,581 | 4,251 | ||||||
| Facilities, overhead and other |
1,386 | 1,002 | ||||||
|
|
|
|
|
|||||
| Total research and development |
41,637 | 18,792 | ||||||
| General and administrative: |
||||||||
| Personnel-related |
3,353 | 1,577 | ||||||
| Third-party service providers |
4,206 | 4,689 | ||||||
| Facilities, overhead and other |
1,034 | 311 | ||||||
|
|
|
|
|
|||||
| Total general and administrative |
8,593 | 6,577 | ||||||
| Total operating expenses |
50,230 | 25,369 | ||||||
|
|
|
|
|
|||||
| Loss from operations |
(50,230 | ) | (25,369 | ) | ||||
|
|
|
|
|
|||||
| Other income (expense), net: |
||||||||
| Interest income |
1,320 | 337 | ||||||
| Interest expense – related party |
— | (327 | ) | |||||
| Other income (expense) |
(2 | ) | (2 | ) | ||||
|
|
|
|
|
|||||
| Total other income (expense), net |
1,318 | 8 | ||||||
|
|
|
|
|
|||||
| Net loss |
$ | (48,912 | ) | $ | (25,361 | ) | ||
|
|
|
|
|
|||||
F-45
Table of Contents
13. Net Loss Per Share Attributable to Common Stockholders
The following table sets forth the computation of basic and diluted net loss per share attributable to common stockholders (in thousands, except share and per share data):
| SIX MONTHS ENDED JUNE 30, | ||||||||
| 2026 | 2025 | |||||||
| Numerator: |
||||||||
| Net loss |
$ | (48,912 | ) | $ | (25,361 | ) | ||
| Deemed dividend to Series C redeemable convertible preferred stockholders |
(9,675 | ) | — | |||||
|
|
|
|
|
|||||
| Net loss attributable to common stockholders |
$ | (58,587 | ) | $ | (25,361 | ) | ||
|
|
|
|
|
|||||
| Denominator: |
||||||||
| Weighted average shares outstanding used to compute basic and diluted net loss per share |
29,329 | 29,329 | ||||||
|
|
|
|
|
|||||
| Net loss per share attributable to common stockholders – basic and diluted: |
$ | (1,997.58 | ) | $ | (864.71 | ) | ||
|
|
|
|
|
|||||
The following outstanding shares of potentially dilutive securities were excluded from the computation of diluted net loss per share for the periods presented because including them would have been anti-dilutive:
| AS OF JUNE 30, | ||||||||
| 2026 | 2025 | |||||||
| Redeemable convertible preferred stock |
39,404,292 | 21,271,235 | ||||||
| Stock options outstanding |
5,701,943 | 1,753,601 | ||||||
|
|
|
|
|
|||||
| Total |
45,106,235 | 23,024,836 | ||||||
|
|
|
|
|
|||||
Electra LLC PI Units are excluded as they are not potentially dilutive securities to the Company.
14. Subsequent events
The Company evaluated subsequent events through August 10, 2026, the date on which these condensed financial statements were available for issuance. No subsequent events have been identified for disclosure to or adjustment in the financial statements.
F-46
Table of Contents
21,666,667 Shares
Common Stock
PROSPECTUS
Jefferies
TD Cowen
Evercore ISI
Cantor
, 2026
Through and including , 2026 (the 25th day after the date of this prospectus), all dealers effecting transactions in these securities, whether or not participating in this offering, may be required to deliver a prospectus. This delivery requirement is in addition to the obligation of dealers to deliver a prospectus when acting as underwriters and with respect to their unsold allotments or subscriptions.
Table of Contents
PART II
INFORMATION NOT REQUIRED IN PROSPECTUS
Item 13. Other Expenses of Issuance and Distribution.
The following table indicates the expenses to be incurred in connection with the offering described in this registration statement, other than underwriting discounts and commissions, all of which will be paid by us. All amounts are estimated except the U.S. Securities and Exchange Commission (SEC) registration fee, the Financial Industry Regulatory Authority, Inc. (FINRA) filing fee, and the exchange listing fee.
| AMOUNT PAID OR TO BE PAID |
||||
| SEC registration fee |
$ | 55,056 | ||
| FINRA filing fee |
60,300 | |||
| Nasdaq Global Select Market listing fee |
325,000 | |||
| Accountants’ fees and expenses |
1,050,000 | |||
| Legal fees and expenses |
3,500,000 | |||
| Transfer agent’s fees and expenses |
10,000 | |||
| Printing and engraving expenses |
450,000 | |||
| Miscellaneous |
199,644 | |||
| Total expenses |
$ | 5,650,000 | ||
Item 14. Indemnification of Directors and Officers.
As permitted by Sections 102 and 145 of the General Corporation Law of the State of Delaware (DGCL), we have adopted provisions in our certificate of incorporation and bylaws that limit or eliminate the personal liability of our directors for a breach of their fiduciary duty of care as a director. The duty of care generally requires that, when acting on behalf of the corporation, directors exercise an informed business judgment based on all material information reasonably available to them. Consequently, a director will not be personally liable to us or our stockholders for monetary damages for breach of fiduciary duty as a director, except for liability for:
| ∎ | any breach of the director’s duty of loyalty to us or our stockholders; |
| ∎ | any act or omission not in good faith or that involves intentional misconduct or a knowing violation of law; |
| ∎ | any act related to unlawful stock repurchases, redemptions, or other distributions or payment of dividends; or |
| ∎ | any transaction from which the director derived an improper personal benefit. |
These limitations of liability do not affect the availability of equitable remedies such as injunctive relief or rescission. Our certificate of incorporation also authorizes us to indemnify our officers, directors, and other agents to the fullest extent permitted under Delaware law.
As permitted by Section 145 of the DGCL, our bylaws provide that:
| ∎ | we may indemnify our directors, officers, employees, and other agents to the fullest extent permitted by the DGCL, subject to limited exceptions; |
| ∎ | we may advance expenses to our directors, officers, and employees in connection with a legal proceeding, subject to limited exceptions; and |
| ∎ | the rights provided in our bylaws are not exclusive. |
Our certificate of incorporation and our bylaws provide for the indemnification provisions described above and elsewhere herein. We have entered or will enter into, and intend to continue to enter into, separate indemnification agreements with our directors and officers that may be broader than the specific indemnification provisions contained in the DGCL. These indemnification agreements generally require us, among other things, to indemnify
II-1
Table of Contents
our officers and directors against certain liabilities that may arise by reason of their status or service as directors or officers, other than liabilities arising from willful misconduct. These indemnification agreements also generally require us to advance any expenses incurred by the directors or officers as a result of any proceeding against them as to which they could be indemnified. These indemnification provisions and the indemnification agreements may be sufficiently broad to permit indemnification of our officers and directors for liabilities, including reimbursement of expenses incurred, arising under the Securities Act of 1933, as amended (Securities Act).
We have purchased and currently intend to maintain insurance on behalf of each and every person who is one of our directors or officers, within the limits and subject to the terms and conditions thereof, against any loss arising from any claim asserted against him or her and incurred by him or her in any such capacity, subject to certain exclusions.
The form of underwriting agreement to be entered into in connection with this initial public offering provides for indemnification by the underwriters of us and our officers and directors who sign this registration statement for specified liabilities, including matters arising under the Securities Act.
Item 15. Recent Sales of Unregistered Securities.
Set forth below is information regarding unregistered securities issued by us since January 1, 2023 to the date of this registration statement. Also included is the consideration received by us for such securities and information relating to the section of the Securities Act, or rule of the SEC, under which exemption from registration was claimed.
| (1) | In March 2025, we issued and sold convertible promissory notes to investors in the aggregate principal amount of $20.0 million. In October 2025, these notes converted into 2,064,955 shares of our Series C redeemable convertible preferred stock. |
| (2) | In June 2025, we issued and sold additional convertible promissory notes to investors in the aggregate principal amount of $20.0 million. In October 2025, these notes converted into 2,015,310 shares of our Series C redeemable convertible preferred stock. |
| (3) | In October 2025, we issued and sold to investors an aggregate of 13,214,580 shares of our Series C redeemable convertible preferred stock, at a purchase price of $10.15 per share for aggregate proceeds of approximately $132.8 million from cash and the conversion of the outstanding convertible promissory notes in paragraphs (1) and (2) above and accrued interest thereon. |
| (4) | In June 2026, we issued and sold to investors an aggregate of 4,918,477 shares of our Series C redeemable convertible preferred stock, at a purchase price of $10.15 per share for aggregate cash proceeds of approximately $49.9 million. |
| (5) | From January 2023 through the date of this registration statement, we granted under our 2022 Stock Plan (2022 Plan) stock options to purchase an aggregate of 5,615,099 shares of our common stock at a weighted-average exercise price of $4.98 per share. From January 2023 through the date of this registration statement, we have issued an aggregate of 94,668 shares of our common stock upon exercise of stock options for an approximate aggregate consideration of $0.5 million. |
The offers, sales, and issuances of the securities described in paragraphs (1) through (4) were deemed to be exempt from registration under the Securities Act in reliance on Section 4(a)(2) of the Securities Act (or Regulation D promulgated thereunder). The recipients of securities in these transactions acquired the securities for investment only and not with a view to or for sale in connection with any distribution thereof and appropriate legends were affixed to the securities issued in such transaction.
Each of the recipients of securities in these transactions was able to bear the investment’s economic risk and had access to the type of information normally provided in a prospectus for a registered securities offering.
The offers, sales, and issuances of the securities described in paragraph (5) were deemed to be exempt from registration under the Securities Act in reliance on Rule 701 in that the transactions were under compensatory benefit plans and contracts relating to compensation as provided under Rule 701 or in reliance on Section 4(a)(2). The recipients of such securities were our employees, directors or bona fide consultants and received the securities under our 2022 Plan.
II-2
Table of Contents
All of the foregoing securities are deemed restricted securities for purposes of the Securities Act. All certificates representing the issued shares of capital stock described in this Item 15 included appropriate legends setting forth that the securities had not been registered and the applicable restrictions on transfer.
Item 16. Exhibits and Financial Statement Schedules.
(a) Exhibits.
II-3
Table of Contents
| EXHIBIT NUMBER |
DESCRIPTION OF EXHIBIT | |
| 23.2 | Consent of Cooley LLP (included in Exhibit 5.1). | |
| 24.1^ | Power of Attorney (included on signature page). | |
| 107 | Filing Fee Table. | |
| ^ | Previously filed |
| + | To be filed by amendment. |
| * | Indicates a management contract or any compensatory plan, contract or arrangement. |
| # | Certain schedules or exhibits to this exhibit have been omitted pursuant to Item 601(a)(5) of Regulation S-K. A copy of any omitted schedule and/or exhibit will be furnished to the SEC upon request. |
(b) Financial Statement Schedules.
All financial statement schedules are omitted because the information required to be set forth therein is not applicable or is shown in the financial statements or the notes thereto.
Item 17. Undertakings.
The undersigned Registrant hereby undertakes to provide to the underwriters at the closing specified in the Underwriting Agreement certificates in such denominations and registered in such names as required by the underwriters to permit prompt delivery to each purchaser.
Insofar as indemnification for liabilities arising under the Securities Act may be permitted to directors, officers and controlling persons of the Registrant pursuant to the foregoing provisions, or otherwise, the Registrant has been advised that in the opinion of the SEC such indemnification is against public policy as expressed in the Securities Act and is, therefore, unenforceable. In the event that a claim for indemnification against such liabilities (other than the payment by the Registrant of expenses incurred or paid by a director, officer or controlling person of the Registrant in the successful defense of any action, suit or proceeding) is asserted by such director, officer, or controlling person in connection with the securities being registered, the Registrant will, unless in the opinion of its counsel the matter has been settled by controlling precedent, submit to a court of appropriate jurisdiction the question whether such indemnification by it is against public policy as expressed in the Securities Act and will be governed by the final adjudication of such issue.
The undersigned Registrant hereby undertakes that:
| (a) | For purposes of determining any liability under the Securities Act, the information omitted from the form of prospectus filed as part of this registration statement in reliance upon Rule 430A and contained in a form of prospectus filed by the Registrant pursuant to Rule 424(b)(1) or (4) or 497(h) under the Securities Act shall be deemed to be part of this registration statement as of the time it was declared effective. |
| (b) | For the purpose of determining any liability under the Securities Act, each post-effective amendment that contains a form of prospectus shall be deemed to be a new registration statement relating to the securities offered therein, and the offering of such securities at that time shall be deemed to be the initial bona fide offering thereof. |
II-4
Table of Contents
Pursuant to the requirements of the Securities Act of 1933, as amended, the Registrant has duly caused this registration statement to be signed on its behalf by the undersigned, thereunto duly authorized, in the City of South San Francisco, California, on September 14, 2026.
| Electra Therapeutics, Inc. | ||
| By: | /s/ Quehuong (Kathy) Dong, Pharm.D., M.B.A. | |
| Quehuong (Kathy) Dong, Pharm.D., M.B.A. | ||
| President and Chief Executive Officer | ||
POWER OF ATTORNEY
Pursuant to the requirements of the Securities Act of 1933, as amended, this registration statement has been signed by the following persons in the capacities and on the dates indicated.
| SIGNATURE |
TITLE |
DATE | ||
| /s/ Quehuong (Kathy) Dong, Pharm.D., M.B.A. Quehuong (Kathy) Dong, Pharm.D., M.B.A. |
President, Chief Executive Officer and Director |
September 14, 2026 | ||
| /s/ Chris Clark, C.F.A. Chris Clark, C.F.A. |
Chief Financial Officer (Principal Financial Officer) |
September 14, 2026 | ||
| /s/ David J. Tucker David J. Tucker |
Senior Vice President, Finance (Principal Accounting Officer) |
September 14, 2026 | ||
| * Nancy Stagliano, Ph.D. |
Chairperson |
September 14, 2026 | ||
| * Matthew Fust, M.B.A. |
Director |
September 14, 2026 | ||
| * Thomas Geninatti, Ph.D. |
Director |
September 14, 2026 | ||
| * Carl L. Gordon, Ph.D., C.F.A. |
Director |
September 14, 2026 | ||
| * Beth Seidenberg, M.D. |
Director |
September 14, 2026 | ||
| *By: | /s/ Chris Clark, C.F.A. | |
| Chris Clark, C.F.A. | ||
| Attorney-in-Fact |
Exhibit 1.1
[ • ] Shares
Electra Therapeutics, Inc.
UNDERWRITING AGREEMENT
[ • ], 2026
Jefferies LLC
TD Securities (USA) LLC
Evercore Group L.L.C.
Cantor Fitzgerald & Co.
As Representatives of the Several Underwriters
c/o Jefferies LLC
520 Madison Avenue
New York, New York 10022
c/o TD Securities (USA) LLC
1 Vanderbilt Avenue
New York, New York 10017
c/o Evercore Group L.L.C.
55 E. 52nd St.
New York, New York 10055
c/o Cantor Fitzgerald & Co.
110 E. 59th St., 6th Floor
New York, New York 10020
Ladies and Gentlemen:
Introductory. Electra Therapeutics, Inc., a Delaware corporation (the “Company”), proposes to issue and sell to the several underwriters named in Schedule A (the “Underwriters”) an aggregate of [•] shares of its common stock, par value $0.0001 per share (the “Shares”). The [•] Shares to be sold by the Company are called the “Firm Shares.” In addition, the Company has granted to the Underwriters an option to purchase up to an additional [•] Shares as provided in Section 2. The additional [•] Shares to be sold by the Company pursuant to such option are collectively called the “Optional Shares.” The Firm Shares and, if and to the extent such option is exercised, the Optional Shares are collectively called the “Offered Shares.” Jefferies LLC (“Jefferies”), TD Securities (USA) LLC (“TD Cowen”), Evercore Group L.L.C. (“Evercore ISI”) and Cantor Fitzgerald & Co. (“Cantor”) have agreed to act as representatives of the several Underwriters (in such capacity, the “Representatives”) in connection with the offering and sale of the Offered Shares. To the extent there are no additional underwriters listed on Schedule A, the term “Representatives” as used herein shall mean you, as Underwriters, and the term “Underwriters” shall mean either the singular or the plural, as the context requires.
The Company has prepared and filed with the Securities and Exchange Commission (the “Commission”) a registration statement on Form S-1, File No. 333-298617 which contains a form of prospectus to be used in connection with the public offering and sale of the Offered Shares. Such registration statement, as amended, including the financial statements, exhibits and schedules thereto, in the form in which it became effective under the Securities Act of 1933, as amended, and the rules and regulations promulgated thereunder (collectively, the “Securities Act”), including any information deemed to be a part thereof at the time of effectiveness pursuant to Rule 430A under the Securities Act, is called the “Registration Statement.” Any registration statement filed by the Company pursuant to Rule 462(b) under the Securities Act in connection with the offer and sale of the Offered Shares is called the “Rule 462(b) Registration Statement,” and from and after the date and time of filing of any such Rule 462(b) Registration Statement the term “Registration Statement” shall include the Rule 462(b) Registration Statement. The prospectus, in the form first used by the Underwriters to confirm sales of the Offered Shares or in the form first made available to the Underwriters by the Company to meet requests of purchasers pursuant to Rule 173 under the Securities Act, is called the “Prospectus.” The preliminary prospectus dated [•], 2026 describing the Offered Shares and the offering thereof is called the “Preliminary Prospectus,” and the Preliminary Prospectus and any other prospectus in preliminary form that describes the Offered Shares and the offering thereof and is used prior to the filing of the Prospectus is called a “preliminary prospectus.” As used herein, “Applicable Time” is [•][a.m.][p.m.] (New York City time) on [•], 2026. As used herein, “free writing prospectus” has the meaning set forth in Rule 405 under the Securities Act, and “Time of Sale Prospectus” means the Preliminary Prospectus together with the free writing prospectuses, if any, identified in Schedule B hereto and the information identified in Schedule D hereto. As used herein, “Road Show” means a “road show” (as defined in Rule 433 under the Securities Act) relating to the offering of the Offered Shares contemplated hereby that is a “written communication” (as defined in Rule 405 under the Securities Act). As used herein, “Section 5(d) Written Communication” means each written communication (within the meaning of Rule 405 under the Securities Act) that is made in reliance on Section 5(d) of the Securities Act by the Company or any person authorized to act on behalf of the Company to one or more potential investors that are qualified institutional buyers (“QIBs”) and/or institutions that are accredited investors (“IAIs”), as such terms are respectively defined in Rule 144A and Rule 501(a) under the Securities Act, to determine whether such investors might have an interest in the offering of the Offered Shares; “Section 5(d) Oral Communication” means each oral communication, if any, made in reliance on Section 5(d) of the Securities Act by the Company or any person authorized to act on behalf of the Company made to one or more QIBs and/or one or more IAIs to determine whether such investors might have an interest in the offering of the Offered Shares; “Marketing Materials” means any materials or information provided to investors by, or with the approval of, the Company in connection with the marketing of the offering of the Offered Shares, including any roadshow or investor presentations made to investors by the Company (whether in person or electronically); and “Permitted Section 5(d) Communication” means the Section 5(d) Written Communication(s) and Marketing Materials listed on Schedule C attached hereto.
All references in this Agreement to (i) the Registration Statement, any preliminary prospectus (including the Preliminary Prospectus), or the Prospectus, or any amendments or supplements to any of the foregoing, or any free writing prospectus, shall include any copy thereof filed with the Commission pursuant to its Electronic Data Gathering, Analysis and Retrieval System (“EDGAR”) and (ii) the Prospectus shall be deemed to include any “electronic Prospectus” provided for use in connection with the offering of the Offered Shares as contemplated by Section 3(n) of this Agreement.
In the event that the Company has only one subsidiary, then all references herein to “subsidiaries” of the Company shall be deemed to refer to such single subsidiary, mutatis mutandis. In the event that the Company has no subsidiaries, all other references to the subsidiaries or any of them in this Agreement shall be disregarded.
2
The Company hereby confirms its agreement with the Underwriters as follows:
Section 1. Representations and Warranties of the Company. The Company hereby represents, warrants and covenants to each Underwriter, as of the date of this Agreement, as of the First Closing Date (as hereinafter defined) and as of each Option Closing Date (as hereinafter defined), if any, as follows:
(a) Compliance with Registration Requirements. The Registration Statement has become effective under the Securities Act. The Company has complied, to the Commission’s satisfaction with all requests of the Commission for additional or supplemental information, if any. No stop order suspending the effectiveness of the Registration Statement is in effect and no proceedings for such purpose have been instituted or are pending or, to the knowledge of the Company, are contemplated or threatened by the Commission.
(b) Disclosure. Each preliminary prospectus and the Prospectus when filed complied in all material respects with the Securities Act and, if filed by electronic transmission pursuant to EDGAR, was identical (except as may be permitted by Regulation S-T under the Securities Act) to the copy thereof delivered to the Underwriters for use in connection with the offer and sale of the Offered Shares. Each of the Registration Statement and any post-effective amendment thereto, at the time it became or becomes effective, complied and will comply in all material respects with the Securities Act and did not and will not contain any untrue statement of a material fact or omit to state a material fact required to be stated therein or necessary to make the statements therein not misleading. As of the Applicable Time, the Time of Sale Prospectus (including any preliminary prospectus wrapper) did not, and at the First Closing Date (as defined in Section 2) and at each applicable Option Closing Date (as defined in Section 2), will not, contain any untrue statement of a material fact or omit to state a material fact necessary to make the statements therein, in the light of the circumstances under which they were made, not misleading. The Prospectus (including any prospectus wrapper), as of its date, did not, and at the First Closing Date and at each applicable Option Closing Date, will not, contain any untrue statement of a material fact or omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances under which they were made, not misleading. The representations and warranties set forth in the three immediately preceding sentences do not apply to statements in or omissions from the Registration Statement or any post-effective amendment thereto, or the Prospectus or the Time of Sale Prospectus, or any amendments or supplements thereto, made in reliance upon and in conformity with written information relating to any Underwriter furnished to the Company in writing by the Representatives expressly for use therein, it being understood and agreed that the only such information consists of the information described in Section 9(b) below. There are no contracts or other documents required to be described in the Time of Sale Prospectus or the Prospectus or to be filed as an exhibit to the Registration Statement which have not been described or filed as required.
(c) Free Writing Prospectuses; Road Show. As of the determination date referenced in Rule 164(h) under the Securities Act, the Company was not, is not or will not be (as applicable) an “ineligible issuer” in connection with the offering of the Offered Shares pursuant to Rules 164, 405 and 433 under the Securities Act. Each free writing prospectus that the Company is required to file pursuant to Rule 433(d) under the Securities Act has been, or will be, filed with the Commission in accordance with the requirements of the Securities Act. Each free writing prospectus that the Company has filed, or is required to file, pursuant to Rule 433(d) under the Securities Act or that was prepared by or on behalf of or used or referred to by the Company complies or will comply in all material respects with the requirements of Rule 433 under the Securities Act, including timely filing with the Commission, retention and legending, as applicable, and each such free writing prospectus, as of its issue date and at all subsequent times through the completion of the public offer and sale of the Offered Shares did not, does not and will not include any information that conflicted, conflicts or will conflict with the information contained in the Registration Statement, the Prospectus or any preliminary prospectus unless such information has been superseded or modified as of such time. Except for the free writing prospectuses, if any, identified in Schedule B, and
3
electronic road shows, if any, furnished to you before first use, the Company has not prepared, used or referred to, and will not, without your prior written consent (which consent shall not be unreasonably withheld, conditioned or delayed), prepare, use or refer to, any free writing prospectus. Each Road Show, when considered together with the Time of Sale Prospectus, did not, as of the Applicable Time, contain any untrue statement of a material fact or omit to state a material fact necessary to make the statements therein, in the light of the circumstances under which they were made, not misleading.
(d) Distribution of Offering Material By the Company. Prior to the later of (i) the expiration or termination of the option granted to the several Underwriters in Section 2, (ii) the completion of the Underwriters’ distribution of the Offered Shares and (iii) the expiration of 25 days after the date of the Prospectus, the Company has not distributed and will not distribute any offering material in connection with the offering and sale of the Offered Shares other than the Registration Statement, the Time of Sale Prospectus, the Prospectus or any free writing prospectus reviewed and consented to by the Representatives, the free writing prospectuses, if any, identified on Schedule B hereto and any Permitted Section 5(d) Communications.
(e) The Underwriting Agreement. This Agreement has been duly authorized, executed and delivered by the Company.
(f) Authorization of the Offered Shares. The Offered Shares have been duly authorized for issuance and sale pursuant to this Agreement and, when issued and delivered by the Company against payment therefor pursuant to this Agreement, will be validly issued, fully paid and nonassessable, and the issuance and sale of the Offered Shares is not subject to any preemptive rights, rights of first refusal or other similar rights to subscribe for or purchase the Offered Shares that have not been duly waived or satisfied.
(g) No Applicable Registration or Other Similar Rights. There are no persons with registration or other similar rights to have any equity or debt securities registered for sale under the Registration Statement or included in the offering contemplated by this Agreement, except for such rights as have been duly waived.
(h) No Material Adverse Change. Except as otherwise disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus, subsequent to the respective dates as of which information is given in the Registration Statement, the Time of Sale Prospectus and the Prospectus: (i) there has been no material adverse change, or any development that would reasonably be expected to result in a material adverse change, in (A) the condition, financial or otherwise, or in the earnings, business, properties, operations, operating results, assets, liabilities or prospects, whether or not arising from transactions in the ordinary course of business, of the Company and its subsidiaries, considered as one entity or (B) the ability of the Company to consummate the transactions contemplated by this Agreement or perform its obligations hereunder (any such change being referred to herein as a “Material Adverse Change”); (ii) the Company and its subsidiaries, considered as one entity, have not incurred any material liability or obligation, indirect, direct or contingent, including without limitation any losses or interference with their business from fire, explosion, flood, earthquakes, accident or other calamity, whether or not covered by insurance, or from any strike, labor dispute or court or governmental action, order or decree, that are material, individually or in the aggregate, to the Company and its subsidiaries, considered as one entity, and have not entered into any material transactions not in the ordinary course of business; and (iii) there has not been any material decrease in the capital stock or any material increase in any short-term or long-term indebtedness of the Company or its subsidiaries and there has been no dividend or distribution of any kind declared, paid or made by the Company or, except for dividends paid to the Company or other subsidiaries, by any of the Company’s subsidiaries on any class of capital stock, or any repurchase or redemption by the Company or any of its subsidiaries of any class of capital stock.
4
(i) Independent Accountants. Ernst & Young LLP, which has expressed its opinion with respect to the financial statements (which term as used in this Agreement includes the related notes thereto) filed with the Commission as a part of the Registration Statement, the Time of Sale Prospectus and the Prospectus, is (i) an independent registered public accounting firm as required by the Securities Act, and the rules of the Public Company Accounting Oversight Board (“PCAOB”), (ii) in compliance with the applicable requirements relating to the qualification of accountants under Rule 2-01 of Regulation S-X under the Securities Act and (iii) a registered public accounting firm as defined by the PCAOB whose registration has not been suspended or revoked and who has not requested such registration to be withdrawn.
(j) Financial Statements. The financial statements filed with the Commission as a part of the Registration Statement, the Time of Sale Prospectus and the Prospectus present fairly, in all material respects, the consolidated financial position of the Company and its subsidiaries as of the dates indicated and the results of their operations, changes in stockholders’ equity and cash flows for the periods specified. Such financial statements have been prepared in conformity with generally accepted accounting principles as applied in the United States (“GAAP”) applied on a consistent basis throughout the periods involved, except as may be expressly stated in the related notes thereto, and except in the case of unaudited financial statements, which are subject to normal and recurring year-end adjustments and do not contain certain footnotes as permitted by the applicable rules of the Commission. No other financial statements or supporting schedules are required to be included in the Registration Statement, the Time of Sale Prospectus or the Prospectus. The financial data set forth in each of the Registration Statement, the Time of Sale Prospectus and the Prospectus under the captions “Prospectus Summary—Summary Financial Data” and “Capitalization” fairly present, in all material respects, the information set forth therein on a basis consistent with that of the audited financial statements contained in the Registration Statement, the Time of Sale Prospectus and the Prospectus. To the Company’s knowledge, no person who has been suspended or barred from being associated with a registered public accounting firm, or who has failed to comply with any sanction pursuant to Rule 5300 promulgated by the PCAOB, has participated in or otherwise aided the preparation of, or audited, the financial statements, supporting schedules or other financial data filed with the Commission as a part of the Registration Statement, the Time of Sale Prospectus and the Prospectus.
(k) Company’s Accounting System. The Company and each of its subsidiaries make and keep books and records that are accurate in all material respects and maintain a system of internal accounting controls designed to provide reasonable assurance that: (i) transactions are executed in accordance with management’s general or specific authorization; (ii) transactions are recorded as necessary to permit preparation of financial statements in conformity with GAAP as applied in the United States and to maintain accountability for assets; (iii) access to assets is permitted only in accordance with management’s general or specific authorization; and (iv) the recorded accountability for assets is compared with existing assets at reasonable intervals and appropriate action is taken with respect to any differences.
(l) Disclosure Controls and Procedures; Deficiencies in or Changes to Internal Control Over Financial Reporting. The Company has established and maintains disclosure controls and procedures (as defined in Rules 13a-15 and 15d-15 under the Securities Exchange Act of 1934, as amended (the “Exchange Act”)), which (i) are designed to ensure that material information relating to the Company, including its consolidated subsidiaries, is made known to the Company’s principal executive officer and its principal financial officer by others within those entities; (ii) have been evaluated by management of the Company for effectiveness as of the end of the Company’s most recent fiscal quarter; and (iii) are effective in all material respects to perform the functions for which they were established. Since the end of the Company’s most recent audited fiscal year, there have been no significant deficiencies or material weaknesses in the Company’s internal control over financial reporting (whether or not remediated) and no change in the Company’s internal control over financial reporting that has materially affected, or is reasonably likely to materially affect, the Company’s internal control over financial reporting. The Company is not aware of any change in its internal control over financial reporting that has occurred during its most recent fiscal quarter that has materially affected, or is reasonably likely to materially affect, the Company’s internal control over financial reporting.
5
(m) Incorporation and Good Standing of the Company. The Company has been duly incorporated and is validly existing as a corporation in good standing under the laws of the jurisdiction of its incorporation and has the corporate power and authority to own, lease and operate its properties and to conduct its business as described in the Registration Statement, the Time of Sale Prospectus and the Prospectus and to enter into and perform its obligations under this Agreement. The Company is duly qualified as a foreign corporation to transact business and is in good standing in the State of California and each other jurisdiction in which such qualification is required, whether by reason of the ownership or leasing of property or the conduct of business, except where the failure to so qualify or to be in good standing would not reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change.
(n) Subsidiaries. The Company has no subsidiaries (as defined in Rule 405 under the Securities Act).
(o) Capitalization and Other Capital Stock Matters. The authorized, issued and outstanding capital stock of the Company is as set forth in the Registration Statement, the Time of Sale Prospectus and the Prospectus under the caption “Capitalization” (other than for subsequent issuances, if any, pursuant to employee benefit plans, upon the exercise of outstanding options or warrants, or pursuant to the automatic conversion of preferred stock of the Company into shares of common stock as a result of the public offering contemplated thereby, in each case as described in the Registration Statement, the Time of Sale Prospectus and the Prospectus). The Shares (including the Offered Shares) conform in all material respects to the description thereof contained in the Time of Sale Prospectus. All of the issued and outstanding Shares have been duly authorized and validly issued, are fully paid and nonassessable and have been issued in compliance with all applicable federal and state securities laws. None of the outstanding Shares were issued in violation of any preemptive rights, rights of first refusal or other similar rights to subscribe for or purchase securities of the Company. There are no authorized or outstanding options, warrants, preemptive rights, rights of first refusal or other rights to purchase, or equity or debt securities convertible into or exchangeable or exercisable for, any capital stock of the Company or any of its subsidiaries other than those described in the Registration Statement, the Time of Sale Prospectus and the Prospectus. The descriptions of the Company’s stock option, stock bonus and other stock plans or arrangements, and the options or other rights granted thereunder, set forth in the Registration Statement, the Time of Sale Prospectus and the Prospectus accurately and fairly present, in all material respects, the information required to be shown with respect to such plans, arrangements, options and rights.
(p) Stock Exchange Listing. The Offered Shares have been approved for listing on the Nasdaq Global Select Market (“Nasdaq”), subject only to official notice of issuance.
(q) Non-Contravention of Existing Instruments; No Further Authorizations or Approvals Required. Neither the Company nor any of its subsidiaries is in violation of its charter or by-laws, partnership agreement or operating agreement or similar organizational documents, as applicable, or is in default (or, with the giving of notice or lapse of time, would be in default) (“Default”) under any indenture, loan, credit agreement, note, lease, license agreement, contract, franchise or other instrument (including, without limitation, any pledge agreement, security agreement, mortgage or other instrument or agreement evidencing, guaranteeing, securing or relating to indebtedness) to which the Company or any of its subsidiaries is a party or by which it or any of them may be bound, or to which any of their respective properties or assets are subject (each, an “Existing Instrument”), except for such Defaults as would not
6
reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change. The Company’s execution, delivery and performance of this Agreement, consummation of the transactions contemplated hereby and by the Registration Statement, the Time of Sale Prospectus and the Prospectus and the issuance and sale of the Offered Shares (including the use of proceeds from the sale of the Offered Shares as described in the Registration Statement, the Time of Sale Prospectus and the Prospectus under the caption “Use of Proceeds”) (i) have been duly authorized by all necessary corporate action and will not result in any violation of the provisions of the charter or by-laws, partnership agreement or operating agreement or similar organizational documents, as applicable, of the Company or any subsidiary, (ii) will not conflict with or constitute a breach of, or Default or a Debt Repayment Triggering Event (as defined below) under, or result in the creation or imposition of any lien, charge or encumbrance upon any property or assets of the Company or any of its subsidiaries pursuant to, or require the consent of any other party to, any Existing Instrument and (iii) will not result in any violation of any law, administrative regulation or administrative or court decree applicable to the Company or any of its subsidiaries, except in the cases of clauses (ii) and (iii) as would not reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change. No consent, approval, authorization or other order of, or registration or filing with, any court or other governmental or regulatory authority or agency, is required for the Company’s execution, delivery and performance of this Agreement and consummation of the transactions contemplated hereby and by the Registration Statement, the Time of Sale Prospectus and the Prospectus, except such as have been obtained or made by the Company and are in full force and effect under the Securities Act and such as may be required under applicable state securities or blue sky laws or by Financial Industry Regulatory Authority, Inc. (“FINRA”). As used herein, a “Debt Repayment Triggering Event” means any event or condition which gives, or with the giving of notice or lapse of time would give, the holder of any note, debenture or other evidence of indebtedness (or any person acting on such holder’s behalf) the right to require the repurchase, redemption or repayment of all or a portion of such indebtedness by the Company or any of its subsidiaries.
(r) Compliance with Laws. The Company and its subsidiaries have been and are in compliance with all applicable laws, rules and regulations, except where failure to be so in compliance would not reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change.
(s) No Material Actions or Proceedings. There is no action, suit, proceeding, inquiry or investigation brought by or before any legal or governmental entity now pending or, to the Company’s knowledge, threatened, against or affecting the Company or any of its subsidiaries, which would reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change. Except as would not reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change, (i) no labor dispute with the employees of the Company or any of its subsidiaries, or with the employees of any principal supplier, manufacturer, customer or contractor of the Company, exists or, (ii) to the Company’s knowledge, is threatened or imminent.
(t) Intellectual Property Rights. The Company owns or has obtained valid, binding, and enforceable licenses or other rights for, the inventions, patent applications, patents, trademarks, trademark applications, trade names, service names, service marks, copyrights and copyrightable works, trade secrets, domain names and other source indicators, technology, software, computer programs, designs, systems, know-how (including other unpatented and/or unpatentable proprietary or confidential information or procedures) and other worldwide intellectual property, industrial property, proprietary rights and similar rights, including any and all registration and applications for registration therefore that are necessary for, used in, or material to the conduct of its business as currently conducted or as currently proposed to be conducted as described in the Registration Statement, the Time of Sale Prospectus and the Prospectus (collectively, “Company Intellectual Property”), provided that nothing in this sentence shall be construed as a representation or warranty regarding the non-infringement, misappropriation, or other violation of the intellectual property rights of any third party. Except as disclosed in the Registration
7
Statement, the Time of Sale Prospectus and the Prospectus, there are no unreleased liens or security interests which have been filed against any of the Company Intellectual Property. The Company Intellectual Property has not been adjudged by a court of competent jurisdiction to be invalid or unenforceable, in whole or in part. There are no third parties who have rights to any Company Intellectual Property, except as disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus as co-owned with or licensed to the Company. To the Company’s knowledge, there is and has been no material infringement, misappropriation, dilution or other violation by third parties of any Company Intellectual Property. To the knowledge of the Company, the Company’s conduct of its respective businesses as now conducted or proposed to be conducted does not infringe, misappropriate, dilute or otherwise violate any valid intellectual property rights of any third party, or will not, upon commercialization of the Company’s product candidates as described in the Registration Statement, the Time of Sale Prospectus and the Prospectus, infringe, misappropriate or otherwise violate any valid intellectual property rights of any third party. Except as disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus, the Company is the sole owner of the Company Intellectual Property owned by or purported to be owned by it and has the valid and enforceable right to use such Company Intellectual Property without any obligation to any third party, including the obligation to obtain consent to sublicense, and without a duty of accounting to co-owner, as applicable. The Company Intellectual Property is subsisting, valid and enforceable. Except as disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus, the Company is not obligated to pay a material royalty, grant a license or option, or provide other material consideration to any third party in connection with the Company Intellectual Property. The Company has taken all reasonable steps necessary to secure assignments to its title, rights and interests in and to the Company Intellectual Property from their employees, consultants, agents and contractors. All employees, consultants, agents and contractors engaged in the development of the intellectual property on behalf of the Company have executed appropriate invention assignment agreements on or about the respective date of hire, in substantially the form made available to the Underwriters and their counsel, whereby such employees, consultants, agents and contractors presently assign all of their right, title and interest in and to such intellectual property to the Company or to a third party who has an obligation to assign, and did assign, such intellectual property to the Company, and to the Company’s knowledge, no such agreement has been breached or violated. Except as disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus, to the Company’s knowledge, no employee of the Company is in or has been in violation of any material term of any employment contract, patent disclosure agreement, invention assignment agreement, non-competition agreement, non-solicitation agreement, nondisclosure agreement or any restrictive covenant to or with a former employer where the basis of such violation relates to such employee’s employment with the Company. The Company has taken reasonable steps in accordance with normal industry practice to maintain the confidentiality of all Company Intellectual Property which derives value from the confidentiality thereof. Except as disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus, there is no pending or, to the Company’s knowledge, threatened or notices of action, suit, proceeding or claim by others: (A) challenging the Company’s rights in or to any Company Intellectual Property; (B) challenging the validity, enforceability or scope of any Company Intellectual Property; or (C) asserting that the Company infringes, misappropriates, dilutes or otherwise violates, or would, upon the manufacturing or commercialization of any product candidates and processes described in the Registration Statement, the Time of Sale Prospectus and the Prospectus as under development, infringe, misappropriate, dilute or otherwise violate, any patent, trademark, trade name, service name, copyright, trade secret or other proprietary rights of others. To the Company’s knowledge, none of the Company Intellectual Property used or held for use by the Company in its business has been obtained or is being used or held for use by the Company in violation of any contractual obligation binding on the Company or in violation of any rights of any third party. To the Company’s knowledge, the Company has complied with the terms of each agreement pursuant to which intellectual property has been licensed to the Company, and all such agreements are in full force and effect. The Company has no reason to believe that the product candidates or processes described in the Registration Statement, the Time of Sale Prospectus
8
and the Prospectus as under development by the Company will not fall within the scope of the claims of one or more patents or patent applications owned by or exclusively licensed to the Company. Except as disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus, no government funding, facilities or resources of a university, college, other educational institution or research center was used in the development of any Company Intellectual Property that is owned or purported to be owned by the Company that would confer any governmental agency or body, university, college, other educational institution or research center any claim or right of ownership to any such Company Intellectual Property.
(u) Patent Rights. To the knowledge of the Company, the patent applications owned by or exclusively licensed to the Company or under which the Company has rights have been duly and properly filed and each issued patent is being diligently maintained and is valid and enforceable, and the patent applications included in the Company Intellectual Property are subsisting, are being diligently prosecuted. To the knowledge of the Company, the parties involved in preparing and prosecuting such applications have complied or are in the process of complying with their duty of candor and disclosure to the U.S. Patent and Trademark Office (the “USPTO”), and all such requirements in the relevant foreign patent authority having similar requirements as the case may be, in connection with such patents and patent applications for which it has filing, prosecution and/or maintenance responsibilities. There is no pending or, to the Company’s knowledge, threatened derivation or interference proceeding pursuant to 35 U.S.C. §135, involving any patents or patent applications within the Company Intellectual Property that are owned or exclusively licensed to the Company.
(v) All Necessary Permits, etc. The Company and its subsidiaries possess such valid and current licenses, certificates, authorizations, approvals, consents or permits required by state, federal or foreign regulatory agencies or bodies to conduct their respective businesses as currently conducted and as described in the Registration Statement, the Time of Sale Prospectus or the Prospectus (“Permits”) except where the failure to possess would not, individually or in the aggregate, reasonably be expected to have a Material Adverse Change. Neither the Company nor any of its subsidiaries is in violation of, or in default under, any of the Permits or has received any notice of proceedings relating to the revocation or modification of, or non-compliance with, any such certificate, authorization or permit, except in each case, where such violation, default, revocation, modification or non-compliance would not, individually or in the aggregate, reasonably be expected to have a Material Adverse Change.
(w) Title to Properties. Except as would not reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change, the Company and its subsidiaries have good and marketable title to all of the personal property and other assets reflected as owned in the financial statements referred to in Section 1(j) above (or elsewhere in the Registration Statement, the Time of Sale Prospectus or the Prospectus), in each case free and clear of any security interests, mortgages, liens, encumbrances, equities, adverse claims and other defects. The real property, improvements, equipment and personal property held under lease by the Company or any of its subsidiaries are held under valid and enforceable leases, with such exceptions as are not material and do not materially interfere with the use made or proposed to be made of such real property, improvements, equipment or personal property by the Company or such subsidiary. The Company and its subsidiaries do not own any real property.
(x) Tax Law Compliance. The Company and its subsidiaries have filed all required federal, state and foreign income and franchise tax returns or have properly requested extensions thereof and have paid all taxes required to be paid by any of them and, if due and payable, any related or similar assessment, fine or penalty levied against any of them except as may be being contested in good faith and by appropriate proceedings or to the extent that failure to file or pay would not reasonably be expected, individually or in the aggregate to result in a Material Adverse Change. The Company has made adequate charges, accruals and reserves in the applicable financial statements referred to in Section 1(j) above in respect of all federal, state and foreign income and franchise taxes for all periods as to which the tax liability of the Company or any of its subsidiaries has not been finally determined.
9
(y) Insurance. Each of the Company and its subsidiaries is insured by recognized, financially sound and reputable institutions with policies in such amounts and with such deductibles and covering such risks as the Company reasonably believes are generally adequate and customary for their businesses including, but not limited to, policies covering real and personal property owned or leased by the Company and its subsidiaries against theft, damage, destruction, acts of vandalism and earthquakes and policies covering the Company and its subsidiaries for product liability claims and clinical trial liability claims. The Company has no reason to believe that it or any of its subsidiaries will not be able (i) to renew its existing insurance coverage as and when such policies expire or (ii) to obtain comparable coverage from similar institutions as may be necessary or appropriate to conduct its business as now conducted and at a cost that would not reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change. Neither the Company nor any of its subsidiaries has been denied any insurance coverage which it has sought or for which it has applied.
(z) Compliance with Environmental Laws. Except as would not reasonably be expected, individually or in the aggregate, to result in a Material Adverse Change: (i) neither the Company nor any of its subsidiaries is in violation of any applicable federal, state, local or foreign statute, law, rule, regulation, ordinance, code, policy or rule of common law or any judicial or administrative interpretation thereof, including any judicial or administrative order, consent, decree or judgment, relating to pollution or protection of human health, the environment (including, without limitation, ambient air, surface water, groundwater, land surface or subsurface strata) or wildlife, including, without limitation, laws and regulations relating to the release or threatened release of chemicals, pollutants, contaminants, wastes, toxic substances, hazardous substances, petroleum or petroleum products (collectively, “Hazardous Materials”) or to the manufacture, processing, distribution, use, treatment, storage, disposal, transport or handling of Hazardous Materials (collectively, “Environmental Laws”); (ii) the Company and its subsidiaries have all permits, authorizations and approvals required under any applicable Environmental Laws and are each in compliance with their requirements; (iii) there are no pending or, to the Company’s knowledge, threatened administrative, regulatory or judicial actions, suits, demands, demand letters, claims, liens, notices of noncompliance or violation, investigation or proceedings relating to any Environmental Law against the Company or any of its subsidiaries; and (iv) to the Company’s knowledge, there are no events or circumstances that might reasonably be expected to form the basis of an order for clean-up or remediation, or an action, suit or proceeding by any private party or governmental body or agency, against or affecting the Company or any of its subsidiaries relating to Hazardous Materials or any Environmental Laws.
(aa) ERISA Compliance. All “employee benefit plans” (as defined under the Employee Retirement Income Security Act of 1974, as amended, and the regulations and published interpretations thereunder (collectively, “ERISA”)) established or maintained by the Company, its subsidiaries or their “ERISA Affiliates” (as defined below) are in compliance in all material respects with ERISA. “ERISA Affiliate” means, with respect to the Company or any of its subsidiaries, any member of any group of organizations described in Sections 414(b), (c), (m) or (o) of the Internal Revenue Code of 1986, as amended, and the regulations and published interpretations thereunder (the “Code”) of which the Company or such subsidiary is a member. To the knowledge of the Company, none of the following events has occurred or is reasonably likely to occur: (i) a failure to maintain and operate any “employee benefit plan” in compliance with its terms and the requirements of any applicable statutes, orders, rules and regulations, including ERISA and the Code; (ii) a non-exempt prohibited transaction, within the meaning of Section 406 of ERISA or Section 4975 of the Code; (iii) a “reportable event,” within the meaning of Section 4043(c) of ERISA, other than those events as to which notice is waived; (iv) a material increase in the aggregate amount of contributions required to be made to any “employee benefit plan” in the current
10
fiscal year of the Company or its subsidiaries compared to the amount of such contributions made in the most recently completed fiscal year of the Company or its subsidiaries; (v) a material increase in the “accumulated post-retirement benefit obligations” (within the meaning of Statement of Financial Accounting Standards 106) of the Company and its subsidiaries compared to the amount of such obligations in the most recently completed fiscal year of the Company and its subsidiaries; or (vi) the filing of a claim by one or more employees or former employees of the Company or any of its subsidiaries related to their employment, that would in each of the clauses (i) – (vi), individually or in the aggregate, reasonably be expected to result in a Material Adverse Change. No “employee benefit plan” established or maintained by the Company, its subsidiaries or any of their ERISA Affiliates, if such “employee benefit plan” were terminated, would have any “amount of unfunded benefit liabilities” (as defined under ERISA). Neither the Company, its subsidiaries nor any of their ERISA Affiliates has incurred or reasonably expects to incur any liability under (i) Title IV of ERISA with respect to termination of, or withdrawal from, any “employee benefit plan” or (ii) Sections 412, 4971, 4975 or 4980B of the Code. Each employee benefit plan established or maintained by the Company, its subsidiaries or any of their ERISA Affiliates that is intended to be qualified under Section 401(a) of the Code is so qualified and, to the knowledge of the Company, nothing has occurred, whether by action or failure to act, which would cause the loss of such qualification. Neither the Company nor any member of its “controlled group” (defined as any entity, whether or not incorporated, that is under common control with the Company or its subsidiaries within the meaning of Section 4001(a)(14) of ERISA or any entity that would be regarded as a single employer with the Company or its subsidiaries under Section 414(b), (c), (m) or (o) of the Code) maintains, or has maintained, a “multiemployer plan” within the meaning of Section 4001(c)(3) of ERISA or a plan (within the meaning of Section 3(3) of ERISA) subject to Section 412 of the Code or Section 302 of Title IV of ERISA.
(bb) Company Not an “Investment Company.” The Company is not, and will not be, either after receipt of payment for the Offered Shares or after the application of the proceeds therefrom as described under “Use of Proceeds” in the Registration Statement, the Time of Sale Prospectus or the Prospectus, required to register as an “investment company” under the Investment Company Act of 1940, as amended (the “Investment Company Act”).
(cc) No Price Stabilization or Manipulation; Compliance with Regulation M. Neither the Company nor any of its subsidiaries has taken, directly or indirectly, without giving effect to any activities by the Underwriters, any action designed to or that would reasonably be expected to cause or result in stabilization or manipulation of the price of the Shares or of any “reference security” (as defined in Rule 100 of Regulation M under the Exchange Act (“Regulation M”)) with respect to the Shares, whether to facilitate the sale or resale of the Offered Shares or otherwise, and has taken no action which would directly or indirectly violate Regulation M.
(dd) Related-Party Transactions. There are no business relationships or related-party transactions involving the Company or any of its subsidiaries or any other person required to be described in the Registration Statement, the Time of Sale Prospectus or the Prospectus that have not been described as required.
(ee) FINRA Matters. All of the information provided to the Underwriters or to counsel for the Underwriters by the Company, its counsel, its officers and directors and, to the Company’s knowledge, the holders of any securities (debt or equity) or options to acquire any securities of the Company in connection with the offering of the Offered Shares is true, complete and correct in all material respects and compliant with FINRA’s rules and any letters, filings or other supplemental information provided to FINRA pursuant to FINRA Rules is true, complete and correct in all material respects.
11
(ff) Parties to Lock-Up Agreements. The Company has furnished to the Underwriters a letter agreement in the form attached hereto as Exhibit A (the “Lock-up Agreement”) from substantially all of the securityholders of the Company. If any additional persons shall become directors or officers (as defined in Rule 16a-1(f) under the Exchange Act) of the Company prior to the end of the Lock-up Period (as defined below), the Company shall cause each such person, prior to or contemporaneously with their appointment or election as a director or officer of the Company, to execute and deliver to the Representatives a Lock-up Agreement.
(gg) Statistical and Market-Related Data. All statistical, demographic and market-related data included in the Registration Statement, the Time of Sale Prospectus or the Prospectus are based on or derived from sources that the Company believes, after reasonable inquiry, to be reliable and accurate in all material respects. To the extent required, the Company has obtained the written consent to the use of such data from such sources.
(hh) Sarbanes-Oxley Act. There is, and has been, no failure on the part of the Company or any of the Company’s directors or officers, in their capacities as such, to comply with any applicable provision of the Sarbanes-Oxley Act of 2002, as amended, and the rules and regulations promulgated in connection therewith, including Section 402 related to loans and Sections 302 and 906 related to certifications.
(ii) No Unlawful Contributions or Other Payments. Neither the Company nor any of its subsidiaries nor, to the Company’s knowledge, any employee or agent of the Company or any subsidiary, has made any contribution or other payment to any official of, or candidate for, any federal, state or foreign office in violation of any applicable law or of the character required to be disclosed in the Registration Statement, the Time of Sale Prospectus or the Prospectus.
(jj) Anti-Corruption and Anti-Bribery Laws. Neither the Company nor any of its subsidiaries nor any director, officer, or employee of the Company or any of its subsidiaries, nor to the knowledge of the Company, any agent, affiliate or other person acting on behalf of the Company or any of its subsidiaries has, in the course of its actions for, or on behalf of, the Company or any of its subsidiaries (i) used any corporate funds for any unlawful contribution, gift, entertainment or other unlawful expenses relating to political activity; (ii) made or taken any act in furtherance of an offer, promise, or authorization of any direct or indirect unlawful payment or benefit to any foreign or domestic government official or employee, including of any government-owned or controlled entity or public international organization, or any political party, party official, or candidate for political office; (iii) violated or is in violation of any provision of the U.S. Foreign Corrupt Practices Act of 1977, as amended (the “FCPA”), the UK Bribery Act 2010, or any other applicable anti-bribery or anti-corruption law; or (iv) made, offered, authorized, requested, or taken an act in furtherance of any unlawful bribe, rebate, payoff, influence payment, kickback or other unlawful payment or benefit. The Company and its subsidiaries and, to the Company’s knowledge, the Company’s affiliates have conducted their respective businesses in compliance with the FCPA and have instituted and maintain policies and procedures designed to ensure, and which are reasonably expected to continue to ensure, continued compliance therewith.
(kk) Money Laundering Laws. The operations of the Company and its subsidiaries are, and have been conducted at all times, in compliance with applicable financial recordkeeping and reporting requirements of the Currency and Foreign Transactions Reporting Act of 1970, as amended, the money laundering statutes of all applicable jurisdictions, the rules and regulations thereunder and any related or similar applicable rules, regulations or guidelines, issued, administered or enforced by any governmental agency (collectively, the “Money Laundering Laws”) and no action, suit or proceeding by or before any court or governmental agency, authority or body or any arbitrator involving the Company or any of its subsidiaries with respect to the Money Laundering Laws is pending or, to the Company’s knowledge, threatened.
12
(ll) Sanctions. Neither the Company nor any of its subsidiaries, directors, officers, or employees, nor, to the Company’s knowledge, any agent, affiliate or other person acting on behalf of the Company or any of its subsidiaries is currently the subject or the target of any U.S. sanctions administered by the Office of Foreign Assets Control of the U.S. Department of the Treasury (“OFAC”) or the U.S. Department of State, the United Nations Security Council, the European Union, His Majesty’s Treasury of the United Kingdom, or other relevant sanctions authority (collectively, “Sanctions”); nor is the Company or any of its subsidiaries located, organized or resident in a country or territory that is the subject or the target of territory-wide or country-wide Sanctions, including, without limitation, the Crimea region of Ukraine, the non-government controlled areas of the Kherson and Zaporizhzhia regions of Ukraine, the so-called Donetsk People’s Republic and the so-called Luhansk People’s Republic or any other region of Ukraine identified pursuant to Executive Order 14065, Cuba, Iran, Syria (prior to July 1, 2025) and North Korea (each a “Sanctioned Country”); and the Company will not directly or indirectly use the proceeds of this offering, or lend, contribute or otherwise make available such proceeds to any subsidiary, or any joint venture partner or other person or entity, for the purpose of financing the activities of or business with any person, or in any country or territory, that at the time of such financing, is the subject or the target of Sanctions or in any other manner that will result in a violation by any person (including any person participating in the transaction whether as underwriter, advisor, investor or otherwise) of applicable Sanctions. Since April 24, 2019, the Company and its subsidiaries have not knowingly engaged in and are not now knowingly engaged in any dealings or transactions with any person that at the time of the dealing or transaction is or was the subject or the target of Sanctions or with any Sanctioned Country.
(mm) Brokers. Except pursuant to this Agreement, there is no broker, finder or other party that is entitled to receive from the Company any brokerage or finder’s fee or other fee or commission as a result of any transactions contemplated by this Agreement.
(nn) Forward-Looking Statements. Each financial or operational projection or other “forward-looking statement” (as defined by Section 27A of the Securities Act or Section 21E of the Exchange Act) contained in the Registration Statement, the Time of Sale Prospectus or the Prospectus (i) was so included by the Company in good faith and with reasonable basis after due consideration by the Company of the underlying assumptions, estimates and other applicable facts and circumstances and (ii) is accompanied by meaningful cautionary statements identifying those factors that could cause actual results to differ materially from those in such forward-looking statement. No such statement was made with the knowledge of an executive officer or director of the Company that it was false or misleading.
(oo) No Outstanding Loans or Other Extensions of Credit. The Company does not have any outstanding extension of credit, in the form of a personal loan, to or for any director or executive officer (or equivalent thereof) of the Company except for such extensions of credit as are expressly permitted by Section 13(k) of the Exchange Act.
(pp) Cybersecurity. The information technology assets and equipment, computers, systems, networks, hardware, software, websites, applications, and databases in the Company and its subsidiaries’ custody or control (collectively, “IT Systems”) are (i) adequate for, and operate and perform in all material respects as required in connection with the operation of the business of the Company and its subsidiaries as currently conducted, and (ii) to the knowledge of the Company, free and clear of all material bugs, errors, defects, Trojan horses, time bombs, malware and other corruptants. The Company and its subsidiaries have implemented and maintained commercially reasonable physical, technical and administrative controls, policies, procedures, and safeguards designed to maintain and protect their material confidential information and the integrity, continuous operation, redundancy and security of all IT Systems and data, including “Personal Data,” used in connection with their businesses (“IT Systems and Data”). “Personal Data” means any clinical trial participant data, personal and/or personally identifiable data or the equivalent under data privacy and security laws. There have been no material breaches, violations, outages or unauthorized uses of or accesses to such IT Systems and Data, except for those that have been remedied without material cost or liability or the duty to notify any other person, nor as of the date hereof are there any incidents under internal review or investigations relating to the same.
13
(qq) Compliance with Data Privacy Laws. The Company and its subsidiaries are presently, and for the past three (3) years have been, in material compliance with all applicable laws or statutes and all judgments, orders, rules and regulations of any court or arbitrator or governmental or regulatory authority, internal policies and contractual obligations each to the extent governing the privacy and security of IT Systems and Personal Data and to the protection of such IT Systems and Personal Data from unauthorized use, access, misappropriation or modification (each a “Data Protection Requirement”). To comply with the Data Protection Requirements, the Company and its subsidiaries have in place, comply with, and take appropriate steps reasonably designed to ensure compliance in all material respects with their policies and procedures governing data privacy and security and the collection, storage, use, disclosure, handling, and analysis of Personal Data (the “Policies”) in material compliance with Data Protection Requirements. The Company and its subsidiaries have made all disclosures to individuals required by applicable privacy and security laws and regulatory rules or requirements, and none of such disclosures made or contained in such disclosures have been inaccurate or in violation of any applicable privacy and security laws and regulatory rules or requirements in any material respect. Neither the Company nor any subsidiary: (i) has received written notice of any actual or potential liability under or relating to, or actual or alleged material violation of, any applicable Data Protection Requirement, and has no knowledge of any event or condition that would reasonably be expected to result in any such notice; or (ii) is a party to any order, decree, or agreement that imposes any obligation or liability by any governmental authority under any applicable Data Protection Requirement. The Company and its subsidiaries since the effective date of the DOJ DSP Rule: (i) are not a “covered person,” (ii) are not a party to and have not engaged in or directed, any “covered data transaction,” including any investments from “covered persons” that do not constitute a “passive investment,” (iii) have not engaged in transactions with the purpose of evading or avoiding the prohibitions in 28 CFR Part 202, “Access to U.S. Sensitive Personal Data and Government-Related Data by Countries of Concern or Covered Persons,” including any amendments thereto and guidance provided thereunder as in effect on the date hereof (the “DOJ DSP Rule”), in each case (i) through (iii) in violation of the DOJ DSP Rule.
(rr) Emerging Growth Company Status. From the time of initial confidential submission of the Registration Statement to the Commission (or, if earlier, the first date on which the Company engaged in any Section 5(d) Written Communication or any Section 5(d) Oral Communication) through the date hereof, the Company has been and is an “emerging growth company,” as defined in Section 2(a) of the Securities Act (an “Emerging Growth Company”).
(ss) Communications. The Company (i) has not alone engaged in communications with potential investors in reliance on Section 5(d) of the Securities Act other than Permitted Section 5(d) Communications or Section 5(d) Oral Communications with the consent of the Representatives with entities that are QIBs or IAIs or those reasonably believed to be QIBs or IAIs and (ii) has not authorized anyone other than the Representatives to engage in such communications; the Company reconfirms that the Representatives have been authorized to act on its behalf in undertaking Marketing Materials, Section 5(d) Oral Communications and Section 5(d) Written Communications; as of the Applicable Time, each Permitted Section 5(d) Communication, when considered together with the Time of Sale Prospectus, did not, as of the Applicable Time, include an untrue statement of a material fact or omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances under which they were made, not misleading; and each Permitted Section 5(d) Communication, if any, does not, as of the date hereof, conflict with the information contained in the Registration Statement, the Preliminary Prospectus and the Prospectus (except where such Permitted Section 5(d) Communication has been superseded by information contained in the Registration Statement, the Preliminary Prospectus and the Prospectus); and the Company has filed publicly on EDGAR at least 15 calendar days prior to any “road show” (as defined in Rule 433 under the Securities Act), any confidentially submitted registration statement and registration statement amendments relating to the offer and sale of the Offered Shares.
14
(tt) Compliance with Health Care Laws. Except as disclosed in the Registration Statement, the Time of Sale Prospectus and the Prospectus, the Company and its subsidiaries: (i) have operated and currently operate their businesses in compliance in all material respects with all applicable Health Care Laws (as defined below) and any other applicable requirements of the Food and Drug Administration (“FDA”), the Department of Health and Human Services and any comparable foreign, state, local or other governmental entity or regulatory authority to which the Company and its subsidiaries are subject (each, an “Applicable Regulatory Authority”) governing the ownership, testing, development, manufacture, packaging, processing, use, distribution, storage, import, export or disposal of any of the Company or its subsidiaries’ product candidates; (ii) have not received any FDA Form-483, written notice of adverse finding, warning letter, untitled letter or other correspondence or written notice from any court or arbitrator or Applicable Regulatory Authority alleging or asserting material non-compliance with (A) any applicable Health Care Laws or (B) any licenses, certificates, approvals, clearances, exemptions, registrations, authorizations, permits and supplements or amendments thereto required by any such applicable Health Care Laws (“Regulatory Authorizations”); (iii) possess all material Regulatory Authorizations required to conduct their respective businesses as currently conducted in full validity and in full force and effect and are not in violation, in any material respect, of any term of any such material Regulatory Authorization; (iv) have not received written notice of any claim, action, suit, proceeding, hearing, enforcement, investigation, arbitration or other action from any Applicable Regulatory Authority or any other third party alleging that any product operation or activity is in material violation of any applicable Health Care Laws or Regulatory Authorizations and have no knowledge that any Applicable Regulatory Authority or any other third party is considering any such claim, litigation, arbitration, action, suit, investigation or proceeding; (v) have not received written notice that any Applicable Regulatory Authority has taken, is taking or intends to take action to limit, suspend, materially modify or revoke any material Regulatory Authorizations and has no knowledge that any Applicable Regulatory Authority is considering such action; (vi) have filed, obtained, maintained or submitted all material reports, documents, forms, notices, applications, records, claims, submissions and supplements or amendments as required by any applicable Health Care Laws or Regulatory Authorizations and that all such reports, documents, forms, notices, applications, records, claims, submissions and supplements or amendments were materially complete and correct on the date filed (or were materially corrected or supplemented by a subsequent submission); (vii) are not a party to or have any ongoing reporting obligations pursuant to any corporate integrity agreements, deferred or non-prosecution agreements, monitoring agreements, consent decrees, settlement orders, plans of correction or similar agreements with or imposed by any Applicable Regulatory Authority; and (viii) along with their employees, officers and directors, and, to the Company’s knowledge, agents, have not been excluded, suspended or debarred from participation in any government health care program or human clinical research or, to the knowledge of the Company, are subject to a governmental inquiry, investigation, proceeding, or other similar action that would reasonably be expected to result in debarment, suspension, or exclusion. The term “Health Care Laws” means Title XVIII of the Social Security Act, 42 U.S.C. §§ 1395-1395hhh (the Medicare statute); Title XIX of the Social Security Act, 42 U.S.C. §§ 1396-1396v (the Medicaid statute); the Federal Anti-Kickback Statute, 42 U.S.C. § 1320a-7b(b); the civil False Claims Act, 31 U.S.C. §§ 3729 et seq.; the criminal False Claims Act 42 U.S.C. 1320a-7b(a); any criminal laws relating to health care fraud and abuse, including but not limited to 18 U.S.C. Sections 286, 287, 1035, 1347, 1349 and the health care fraud criminal provisions under the Health Insurance Portability and Accountability Act of 1996, 42 U.S.C. §§ 1320d et seq., (“HIPAA”); the Civil Monetary Penalties Law, 42 U.S.C. §§ 1320a-7a and 1320a-7b; the Physician Payments Sunshine Act, 42 U.S.C. § 1320a-7h; the Exclusions Law, 42 U.S.C. § 1320a-7; HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act, 42 U.S.C. §§ 17921 et seq.; the Federal Food, Drug, and Cosmetic Act, 21 U.S.C. §§ 301 et seq.; any other law pertaining to or governing a government health care program; and any similar foreign, federal, state and local laws, in each case as amended, together with the regulations promulgated pursuant to any such laws.
15
(uu) Clinical Data and Regulatory Compliance. All clinical and pre-clinical studies and trials conducted by or on behalf of or sponsored by the Company or any of its subsidiaries, or in which the Company or any of its subsidiaries have participated with respect to the Company’s or any of its subsidiaries’ product candidates, including without limitation any such studies and trials that are described in the Registration Statement, the Time of Sale Prospectus and the Prospectus, or the results of which are referred to in the Registration Statement, the Time of Sale Prospectus and the Prospectus, as applicable (collectively, the “Company Trials”), were, and if still pending are, being conducted in all material respects in accordance with all applicable Health Care Laws and any applicable rules, regulations and policies of the jurisdiction in which such trials and studies were or are being conducted; the descriptions in the Registration Statement, the Time of Sale Prospectus and the Prospectus of the results of any Company Trials are accurate descriptions in all material respects and the Company has no knowledge of any other studies or trials not described in the Registration Statement, the Time of Sale Prospectus and the Prospectus, the results of which are inconsistent with or call into question the results described or referred to in the Registration Statement, the Time of Sale Prospectus and the Prospectus; neither the Company nor any of its subsidiaries have received any written notices, correspondence or other communications from any Applicable Regulatory Authority or any other governmental entity or institutional review board requiring or threatening the termination, material modification or suspension of any Company Trials, other than ordinary course communications with respect to modifications in connection with the design and implementation of such Company Trials, and, to the Company’s knowledge, there are no reasonable grounds for the same. No investigational new drug application or comparable submission filed by or on behalf of the Company or its subsidiaries with the FDA or other Applicable Regulatory Authority has been terminated or suspended by the FDA or such other Applicable Regulatory Authority. To the Company’s knowledge, none of the Company Trials involved any investigator who has been disqualified as a clinical investigator or has been found by the FDA or any other Applicable Regulatory Authority to have engaged in scientific misconduct.
(vv) No Contract Terminations. Neither the Company nor any of its subsidiaries has sent or received any communication regarding termination of, or intent not to renew, any of the contracts or agreements referred to or described in any preliminary prospectus, the Prospectus or any free writing prospectus, or referred to or described in, or filed as an exhibit to, the Registration Statement, and no such termination or non-renewal has been threatened by the Company or any of its subsidiaries or, to the Company’s knowledge, any other party to any such contract or agreement, which threat of termination or non-renewal has not been rescinded as of the date hereof.
(ww) Outbound Investment Security Program. The Company is not a “covered foreign person,” as that term is defined in 31 C.F.R. § 850.209. The Company does not currently engage, or have plans to engage, directly or indirectly, in a “covered activity,” as that term is defined in 31 C.F.R. § 850.208 (“Covered Activity”). The Company does not have any joint ventures that engage in or plan to engage in any Covered Activity. The Company also does not, directly or indirectly, hold a board seat on, have a voting or equity interest in, or have any contractual power to direct or cause the direction of the management or policies of any person or persons that engages or plans to engage in any Covered Activity.
Any certificate signed by any officer of the Company or any of its subsidiaries and delivered to any Underwriter or to counsel for the Underwriters in connection with the offering, or the purchase and sale, of the Offered Shares shall be deemed a representation and warranty by the Company to each Underwriter as to the matters covered thereby.
16
The Company has a reasonable basis for making each of the representations set forth in this Section 1. The Company acknowledges that the Underwriters and, for purposes of the opinions to be delivered pursuant to Section 6 hereof, counsel to the Company and counsel to the Underwriters, will rely upon the accuracy and truthfulness of the foregoing representations and hereby consents to such reliance.
Section 2. Purchase, Sale and Delivery of the Offered Shares.
(a) The Firm Shares. Upon the terms herein set forth, the Company agrees to issue and sell to the several Underwriters an aggregate of [•] Firm Shares. On the basis of the representations, warranties and agreements herein contained, and upon the terms but subject to the conditions herein set forth, the Underwriters agree, severally and not jointly, to purchase from the Company the respective number of Firm Shares set forth opposite their names on Schedule A. The purchase price per Firm Share to be paid by the several Underwriters to the Company shall be $[•] per share.
(b) The First Closing Date. Delivery of the Firm Shares to be purchased by the Underwriters and payment therefor shall be made at the offices of Latham & Watkins LLP (or such other place as may be agreed to by the Company and the Representatives) at 9:00 a.m. New York City time, on [•], 2026, or such other time and date not later than 1:30 p.m. New York City time, on [•], 2026 as the Representatives shall designate by notice to the Company (the time and date of such closing are called the “First Closing Date”). The Company hereby acknowledges that circumstances under which the Representatives may provide notice to postpone the First Closing Date as originally scheduled include, but are not limited to, any determination by the Company or the Representatives to recirculate to the public copies of an amended or supplemented Prospectus or a delay as contemplated by the provisions of Section 11.
(c) The Optional Shares; Option Closing Date. In addition, on the basis of the representations, warranties and agreements herein contained, and upon the terms but subject to the conditions herein set forth, the Company hereby grants an option to the several Underwriters to purchase, severally and not jointly, up to an aggregate of [•] Optional Shares from the Company at the purchase price per share to be paid by the Underwriters for the Firm Shares. The option granted hereunder may be exercised at any time and from time to time in whole or in part upon notice by the Representatives to the Company, which notice may be given at any time within 30 days from the date of this Agreement. Such notice shall set forth (i) the aggregate number of Optional Shares as to which the Underwriters are exercising the option and (ii) the time, date and place at which the Optional Shares will be delivered (which time and date may be simultaneous with, but not earlier than, the First Closing Date; and in the event that such time and date are simultaneous with the First Closing Date, the term “First Closing Date” shall refer to the time and date of delivery of the Firm Shares and such Optional Shares). Any such time and date of delivery, if subsequent to the First Closing Date, is called an “Option Closing Date,” and shall be determined by the Representatives and shall not be earlier than one day or later than five full business days after delivery of such notice of exercise. If any Optional Shares are to be purchased, each Underwriter agrees, severally and not jointly, to purchase the number of Optional Shares (subject to such adjustments to eliminate fractional shares as the Representatives may determine) that bears the same proportion to the total number of Optional Shares to be purchased as the number of Firm Shares set forth on Schedule A opposite the name of such Underwriter bears to the total number of Firm Shares. The Representatives may cancel the option at any time prior to its expiration by giving written notice of such cancellation to the Company.
(d) Public Offering of the Offered Shares. The Representatives hereby advise the Company that the Underwriters intend to offer for sale to the public, initially on the terms set forth in the Registration Statement, the Time of Sale Prospectus and the Prospectus, their respective portions of the Offered Shares as soon after this Agreement has been executed and the Registration Statement has been declared effective as the Representatives, in their sole judgment, have determined is advisable and practicable.
17
(e) Payment for the Offered Shares. (i) Payment for the Firm Shares shall be made at the First Closing Date (and, if applicable, payment for the Optional Shares shall be made at the First Closing Date or the applicable Option Closing Date, as the case may be) by wire transfer of immediately available funds to the order of the Company.
(ii) It is understood that the Representatives have been authorized, for their own account and the accounts of the several Underwriters, to accept delivery of and receipt for, and make payment of the purchase price for, the Firm Shares and any Optional Shares the Underwriters have agreed to purchase. Each of Jefferies, TD Cowen, Evercore ISI and Cantor, individually and not as the Representatives of the Underwriters, may (but shall not be obligated to) make payment for any Offered Shares to be purchased by any Underwriter whose funds shall not have been received by the Representatives by the First Closing Date or the applicable Option Closing Date, as the case may be, for the account of such Underwriter, but any such payment shall not relieve such Underwriter from any of its obligations under this Agreement.
(f) Delivery of the Offered Shares. The Company shall deliver, or cause to be delivered, through the facilities of The Depository Trust Company (“DTC”), to the Representatives for the accounts of the several Underwriters the Firm Shares at the First Closing Date, against release of a wire transfer of immediately available funds for the amount of the purchase price therefor. The Company shall also deliver, or cause to be delivered to the Representatives for the accounts of the several Underwriters, the Optional Shares the Underwriters have agreed to purchase at the First Closing Date or the applicable Option Closing Date, as the case may be, against the release of a wire transfer of immediately available funds for the amount of the purchase price therefor. If Jefferies so elects, delivery of the Offered Shares may be made by credit to the accounts designated by Jefferies through DTC’s full fast transfer or Deposit/Withdrawal at Custodian programs. If Jefferies so elects, the Offered Shares shall be registered in such names and denominations as the Representatives shall have requested at least one business day prior to the First Closing Date (or the applicable Option Closing Date, as the case may be) and shall be made available for inspection on the business day preceding the First Closing Date (or the applicable Option Closing Date, as the case may be) at a location in New York City as the Representatives may designate. Time shall be of the essence, and delivery at the time and place specified in this Agreement is a further condition to the obligations of the Underwriters.
Section 3. Additional Covenants of the Company. The Company further covenants and agrees with each Underwriter as follows:
(a) Delivery of Registration Statement, Time of Sale Prospectus and Prospectus. The Company shall furnish, upon request, to the Representatives in New York City, without charge, prior to 10:00 a.m. New York City time on the business day next following the date of this Agreement and during the period when a prospectus relating to the Offered Shares is required by the Securities Act to be delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule) in connection with sales of the Offered Shares, as many copies of the Time of Sale Prospectus, the Prospectus and any supplements and amendments thereto or to the Registration Statement as the Representatives may reasonably request.
(b) Representatives’ Review of Proposed Amendments and Supplements. During the period when a prospectus relating to the Offered Shares is required by the Securities Act to be delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule), the Company (i) will furnish to the Representatives for review, a reasonable period of time prior to the proposed time of filing of any proposed amendment or supplement to the Registration Statement, a copy of each such amendment or supplement and (ii) will not amend or supplement the Registration Statement without the Representatives’ prior written consent (which consent shall not be unreasonably withheld,
18
conditioned or delayed). Prior to amending or supplementing any preliminary prospectus, the Time of Sale Prospectus or the Prospectus, the Company shall furnish to the Representatives for review, a reasonable amount of time prior to the time of filing or use of the proposed amendment or supplement, a copy of each such proposed amendment or supplement. The Company shall not file or use any such proposed amendment or supplement without the Representatives’ prior written consent (which consent shall not be unreasonably withheld, conditioned or delayed). The Company shall file with the Commission within the applicable period specified in Rule 424(b) under the Securities Act any prospectus required to be filed pursuant to such Rule.
(c) Free Writing Prospectuses. The Company shall furnish to the Representatives for review, a reasonable amount of time prior to the proposed time of filing or use thereof, a copy of each proposed free writing prospectus or any amendment or supplement thereto prepared by or on behalf of, used by, or referred to by the Company, and the Company shall not file, use or refer to any proposed free writing prospectus or any amendment or supplement thereto without the Representatives’ prior written consent (which consent shall not be unreasonably withheld, conditioned or delayed). The Company shall furnish to each Underwriter, without charge, as many copies of any free writing prospectus prepared by or on behalf of, used by or referred to by the Company as such Underwriter may reasonably request. If at any time when a prospectus is required by the Securities Act to be delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule) in connection with sales of the Offered Shares (but in any event if at any time through and including the First Closing Date) there occurred or occurs an event or development as a result of which any free writing prospectus prepared by or on behalf of, used by, or referred to by the Company conflicted or would conflict with the information contained in the Registration Statement or included or would include an untrue statement of a material fact or omitted or would omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances prevailing at such time, not misleading, the Company shall promptly amend or supplement such free writing prospectus to eliminate or correct such conflict or so that the statements in such free writing prospectus as so amended or supplemented will not include an untrue statement of a material fact or omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances prevailing at such time, not misleading, as the case may be; provided, however, that prior to amending or supplementing any such free writing prospectus, the Company shall furnish to the Representatives for review, a reasonable amount of time prior to the proposed time of filing or use thereof, a copy of such proposed amended or supplemented free writing prospectus, and the Company shall not file, use or refer to any such amended or supplemented free writing prospectus without the Representatives’ prior written consent (which consent shall not be unreasonably withheld, conditioned or delayed).
(d) Filing of Underwriter Free Writing Prospectuses. The Company shall not take any action that would result in an Underwriter or the Company being required to file with the Commission pursuant to Rule 433(d) under the Securities Act a free writing prospectus prepared by or on behalf of such Underwriter that such Underwriter otherwise would not have been required to file thereunder.
(e) Amendments and Supplements to Time of Sale Prospectus. If the Time of Sale Prospectus is being used to solicit offers to buy the Offered Shares at a time when the Prospectus is not yet available to prospective purchasers, and any event shall occur or condition exist as a result of which it is necessary to amend or supplement the Time of Sale Prospectus so that the Time of Sale Prospectus does not include an untrue statement of a material fact or omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances when delivered to a prospective purchaser, not misleading, or if any event shall occur or condition exist as a result of which the Time of Sale Prospectus conflicts with the information contained in the Registration Statement, or if, in the opinion of counsel for the Underwriters, it is necessary to amend or supplement the Time of Sale Prospectus to comply with applicable law, the Company shall (subject to Section 3(b) and Section 3(c) hereof) promptly prepare, file
19
with the Commission and furnish, at its own expense, to the Underwriters and to any dealer upon request, either amendments or supplements to the Time of Sale Prospectus so that the statements in the Time of Sale Prospectus as so amended or supplemented will not include an untrue statement of a material fact or omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances when delivered to a prospective purchaser, not misleading or so that the Time of Sale Prospectus, as amended or supplemented, will no longer conflict with the information contained in the Registration Statement, or so that the Time of Sale Prospectus, as amended or supplemented, will comply with applicable law.
(f) Certain Notifications and Required Actions. After the date of this Agreement and until such time as the Underwriters are no longer required to deliver a Prospectus in order to confirm sales of the Offered Shares, the Company shall promptly advise the Representatives in writing (which may be by electronic mail) of: (i) the receipt of any comments of, or requests for additional or supplemental information from, the Commission relating to the Registration Statement received by the Company; (ii) the time and date of any filing of any post-effective amendment to the Registration Statement or any amendment or supplement to any preliminary prospectus, the Time of Sale Prospectus, any free writing prospectus or the Prospectus; (iii) the time and date that any post-effective amendment to the Registration Statement becomes effective; and (iv) the issuance by the Commission of any stop order suspending the effectiveness of the Registration Statement or any post-effective amendment thereto or any amendment or supplement to any preliminary prospectus, the Time of Sale Prospectus or the Prospectus or of any order preventing or suspending the use of any preliminary prospectus, the Time of Sale Prospectus, any free writing prospectus or the Prospectus, or, if the Company gains knowledge of any such proceeding, of any proceedings to remove, suspend or terminate from listing or quotation the Shares from any securities exchange upon which they are listed for trading or included or designated for quotation, or of the threatening or initiation of any proceedings for any of such purposes. If the Commission shall enter any such stop order at any time, the Company will use its best efforts to obtain the lifting of such order as soon as practicable. Additionally, the Company agrees that it shall comply with all applicable provisions of Rule 424(b), Rule 433 and Rule 430A under the Securities Act and will use its reasonable efforts to confirm that any filings made by the Company under Rule 424(b) or Rule 433 were received in a timely manner by the Commission.
(g) Amendments and Supplements to the Prospectus and Other Securities Act Matters. If any event shall occur or condition exist as a result of which it is necessary to amend or supplement the Prospectus so that the Prospectus does not include an untrue statement of a material fact or omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances when the Prospectus is delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule) to a purchaser, not misleading, or if in the opinion of the Representatives or counsel for the Underwriters it is otherwise necessary to amend or supplement the Prospectus to comply with applicable law, the Company agrees (subject to Section 3(b) and Section 3(c) hereof) to promptly prepare, file with the Commission and furnish, at its own expense, to the Underwriters and to any dealer upon request, amendments or supplements to the Prospectus so that the statements in the Prospectus as so amended or supplemented will not include an untrue statement of a material fact or omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances when the Prospectus is delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule) to a purchaser, not misleading or so that the Prospectus, as amended or supplemented, will comply with applicable law. Neither the Representatives’ consent to, nor delivery of, any such amendment or supplement shall constitute a waiver of any of the Company’s obligations under Section 3(b) or Section 3(c).
20
(h) Blue Sky Compliance. The Company shall cooperate with the Representatives and counsel for the Underwriters to qualify or register the Offered Shares for sale under (or obtain exemptions from the application of) the state securities or blue sky laws (or other foreign laws) of those jurisdictions as may be reasonably designated by the Representatives, shall comply with such laws and shall continue such qualifications, registrations and exemptions in effect so long as required for the distribution of the Offered Shares. The Company shall not be required to qualify as a foreign corporation or to take any action that would subject it to general service of process in any such jurisdiction where it is not presently qualified or where it would be subject to taxation as a foreign corporation. The Company will advise the Representatives promptly of the suspension of the qualification or registration of (or any such exemption relating to) the Offered Shares for offering, sale or trading in any jurisdiction or, if the Company gains knowledge of any such proceeding, any initiation or threat of any proceeding for any such purpose, and in the event of the issuance of any order suspending such qualification, registration or exemption, the Company shall use its reasonable best efforts to obtain the withdrawal thereof as soon as practicable.
(i) Use of Proceeds. The Company shall apply the net proceeds from the sale of the Offered Shares sold by it in all material respects in the manner described under the caption “Use of Proceeds” in the Registration Statement, the Time of Sale Prospectus and the Prospectus.
(j) Transfer Agent. The Company shall engage and maintain, at its expense, a registrar and transfer agent for the Shares.
(k) Earnings Statement. The Company will make generally available to its security holders and to the Representatives as soon as practicable an earnings statement (which need not be audited) covering a period of at least twelve months beginning with the first fiscal quarter of the Company commencing after the date of this Agreement that will satisfy the provisions of Section 11(a) of the Securities Act and the rules and regulations of the Commission thereunder; provided, however, that the requirements of this Section 3(k) shall be deemed satisfied to the extent such statement is available on EDGAR.
(l) Continued Compliance with Securities Laws. The Company will comply with the Securities Act and the Exchange Act so as to permit the completion of the distribution of the Offered Shares as contemplated by this Agreement, the Registration Statement, the Time of Sale Prospectus and the Prospectus. Without limiting the generality of the foregoing, the Company will, during the period when a prospectus relating to the Offered Shares is required by the Securities Act to be delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule), file on a timely basis with the Commission and Nasdaq all reports and documents required to be filed under the Exchange Act. Additionally, the Company shall report the use of proceeds from the issuance of the Offered Shares as may be required under Rule 463 under the Securities Act.
(m) Listing. The Company will use its best efforts to list, subject to notice of issuance, the Offered Shares on Nasdaq.
(n) Company to Provide Copy of the Prospectus in Form That May be Downloaded from the Internet. If requested by the Representatives, the Company shall cause to be prepared and delivered, at its expense, within one business day from the effective date of this Agreement, to the Representatives an “electronic Prospectus” to be used by the Underwriters in connection with the offering and sale of the Offered Shares. As used herein, the term “electronic Prospectus” means a form of Prospectus, and any amendment or supplement thereto, that meets each of the following conditions: (i) it shall be encoded in an electronic format, satisfactory to the Representatives, that may be transmitted electronically by the Representatives and the other Underwriters to offerees and purchasers of the Offered Shares; (ii) it shall disclose the same information as the paper Prospectus, except to the extent that graphic and image material cannot be disseminated electronically, in which case such graphic and image material shall be replaced in the electronic Prospectus with a fair and accurate narrative description or tabular representation of such
21
material, as appropriate; and (iii) it shall be in or convertible into a paper format or an electronic format, satisfactory to the Representatives, that will allow investors to store and have continuously ready access to the Prospectus at any future time, without charge to investors (other than any fee charged for subscription to the Internet as a whole and for on-line time). The Company hereby confirms that it has included or will include in the Prospectus filed pursuant to EDGAR or otherwise with the Commission and in the Registration Statement at the time it was declared effective an undertaking that, upon receipt of a request by an investor or his or her representative, the Company shall transmit or cause to be transmitted promptly, without charge, a paper copy of the Prospectus.
(o) Agreement Not to Offer or Sell Additional Shares. During the period commencing on and including the date hereof and continuing through and including the 180th day following the date of the Prospectus (such period being referred to herein as the “Lock-up Period”), the Company will not, without the prior written consent of Jefferies, TD Cowen and Evercore ISI (which consent may be withheld in their sole discretion), directly or indirectly: (i) sell, offer to sell, contract to sell or lend any Shares or Related Securities (as defined below); (ii) effect any short sale, or establish or increase any “put equivalent position” (as defined in Rule 16a-1(h) under the Exchange Act) or liquidate or decrease any “call equivalent position” (as defined in Rule 16a-1(b) under the Exchange Act) of any Shares or Related Securities; (iii) pledge, hypothecate or grant any security interest in any Shares or Related Securities; (iv) in any other way transfer or dispose of any Shares or Related Securities; (v) enter into any swap, hedge or similar arrangement or agreement that transfers, in whole or in part, the economic risk of ownership of any Shares or Related Securities, regardless of whether any such transaction is to be settled in securities, in cash or otherwise; (vi) announce the offering of any Shares or Related Securities; (vii) submit or file any registration statement under the Securities Act in respect of any Shares or Related Securities (other than as contemplated by this Agreement with respect to the Offered Shares); (viii) effect a reverse stock split, recapitalization, share consolidation, reclassification or similar transaction affecting the outstanding Shares; or (ix) publicly announce the intention to do any of the foregoing; provided, however, that the Company may (A) effect the transactions contemplated hereby, (B) issue Shares or Related Securities, or issue Shares upon exercise of options, pursuant to any stock option, stock bonus, employee stock purchase plan, or other equity plan or arrangement described in the Registration Statement, the Time of Sale Prospectus and the Prospectus, but only if the holders of such Shares or options agree in writing with the Underwriters not to sell, offer, dispose of or otherwise transfer any such Shares or options during such Lock-up Period without the prior written consent of Jefferies, TD Cowen and Evercore ISI (which consent may be withheld in their sole discretion), (C) issue Shares or Related Securities to any non-employee director pursuant to any non-employee director compensation policy or program described in the Registration Statement, the Time of Sale Prospectus and the Prospectus, (D) issue Shares pursuant to the exercise or settlement of Related Securities or upon the conversion of convertible securities outstanding on the date hereof that are described in the Registration Statement, the Time of Sale Prospectus and the Prospectus, provided the recipient to any Shares or Related Securities (solely to the extent any such Related Securities are or may become exercisable or settleable for Shares during the Lock-up Period) pursuant to subsections (C) and (D), as applicable, executes a Lock-up Agreement in substantially the form attached as Exhibit A, (E) file one or more registration statements on Form S-8 or a successor form thereto to register Shares or Related Securities issued or issuable pursuant to the terms of a stock option, stock bonus or other equity plan or arrangement described in (B) or (C) above; and (F) issue Shares or any Related Securities issued in connection with any merger, joint venture, strategic alliance, commercial or other strategic or collaborative relationship or the acquisition or license by the Company of the securities, businesses, property or other assets of another person or entity or pursuant to any employment benefit plan assumed by the Company in connection with any such merger or acquisition, provided that, in the case of clause (F), (i) the aggregate number of Shares that the Company may sell or issue or agree to sell or issue pursuant to clause (F) shall not exceed 5.0% of the total number of Shares issued and outstanding immediately following the issuance of the Offered Shares and (ii) the recipients thereof provide to the Representatives a signed Lock-up Agreement in substantially the form attached as Exhibit A. For purposes of the foregoing, “Related Securities” shall mean any options or warrants or other rights to acquire Shares or any securities exchangeable or exercisable for or convertible into Shares, or to acquire other securities or rights ultimately exchangeable or exercisable for, or convertible into, Shares.
22
(p) Future Reports to the Representatives. During the period of five years hereafter, the Company will furnish to the Representatives, c/o Jefferies, at 520 Madison Avenue, New York, New York 10022, Attention: General Counsel, c/o TD Cowen, 1 Vanderbilt Avenue, New York, New York 10017, Attention: Head of Equity Capital Markets; General Counsel, Email: ciblegal@tdsecurities.com, c/o Evercore ISI, 55 East 52nd Street, 35th Floor, New York, New York 10055, Attention: ECM General Counsel and c/o Cantor Fitzgerald & Co., 110 East 59th Street, 6th Floor, New York, New York 10022, Attention: General Counsel: (i) as soon as practicable after the end of each fiscal year, copies of the Annual Report of the Company containing the balance sheet of the Company as of the close of such fiscal year and statements of income, stockholders’ equity and cash flows for the year then ended and the opinion thereon of the Company’s independent public or certified public accountants; (ii) as soon as practicable after the filing thereof, copies of each proxy statement, Annual Report on Form 10-K, Quarterly Report on Form 10-Q, Current Report on Form 8-K or other report filed by the Company with the Commission or any securities exchange; and (iii) as soon as available, copies of any report or communication of the Company furnished or made available generally to holders of its capital stock; provided, however, that the requirements of this Section 3(p) shall be satisfied to the extent that such reports, statement, communications, financial statements or other documents are available on EDGAR.
(q) Investment Limitation. The Company shall not invest or otherwise use the proceeds received by the Company from its sale of the Offered Shares in such a manner as would require the Company or any of its subsidiaries to register as an investment company under the Investment Company Act.
(r) No Stabilization or Manipulation; Compliance with Regulation M. The Company will not take, and will ensure that no affiliate of the Company will take, directly or indirectly, without giving effect to activities by the Underwriters, any action designed to or that would reasonably be expected to cause or result in stabilization or manipulation of the price of the Shares or any reference security with respect to the Shares, whether to facilitate the sale or resale of the Offered Shares or otherwise, and the Company will, and shall cause each of its affiliates to, comply with all applicable provisions of Regulation M.
(s) Enforce Lock-Up Agreements. During the Lock-up Period, the Company will enforce all agreements between the Company and any of its securityholders that restrict or prohibit, expressly or in operation, the offer, sale or transfer of Shares or Related Securities or any of the other actions restricted or prohibited under the terms of the form of Lock-up Agreement. In addition, the Company will direct the transfer agent to place stop transfer restrictions upon any such securities of the Company that are bound by such “lock-up” agreements for the duration of the periods contemplated in such agreements, including, without limitation, “lock-up” agreements entered into by the Company’s officers, directors and securityholders pursuant to Section 6(i) hereof.
(t) Company to Provide Interim Financial Statements. Prior to the First Closing Date and each applicable Option Closing Date, the Company will furnish the Underwriters, as soon as reasonably practicable, a copy of any unaudited interim financial statements of the Company for any period subsequent to the period covered by the most recent financial statements appearing in the Registration Statement and the Prospectus; provided that the requirements of this Section 3(t) shall be deemed satisfied to the extent such financial statements are available on EDGAR.
23
(u) Amendments and Supplements to Permitted Section 5(d) Communications. If at any time following the distribution of any Permitted Section 5(d) Communication, during the period when a prospectus relating to the Offered Shares is required by the Securities Act to be delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule), there occurred or occurs an event or development as a result of which such Permitted Section 5(d) Communication included or would include an untrue statement of a material fact or omitted or would omit to state a material fact necessary in order to make the statements therein, in the light of the circumstances existing at that subsequent time, not misleading, the Company will promptly notify the Representatives and, upon the reasonable request of the Representatives, will promptly amend or supplement, at its own expense, such Permitted Section 5(d) Communication to eliminate or correct such untrue statement or omission.
(v) Emerging Growth Company Status. The Company will promptly notify the Representatives if the Company ceases to be an Emerging Growth Company at any time prior to the later of (i) the time when a prospectus relating to the Offered Shares is not required by the Securities Act to be delivered (whether physically or through compliance with Rule 172 under the Securities Act or any similar rule) and (ii) the expiration of the Lock-up Period (as defined herein).
(w) Announcement Regarding Lock-ups. The Company agrees to announce the applicable Representatives’ intention to release any director or “officer” (within the meaning of Rule 16a-1(f) under the Exchange Act) of the Company from any of the restrictions imposed by any Lock-Up Agreement, by issuing, through a major news service, a press release in form and substance satisfactory to such Representatives or, if consented to by such Representatives, in a registration statement that is publicly filed in connection with a secondary offering of the Company’s shares promptly following the Company’s receipt of any notification from such Representatives in which such intention is indicated, but in any case not later than the close of the third business day prior to the date on which such release or waiver is to become effective; provided, however, that nothing shall prevent the applicable Representatives, on behalf of the Underwriters, from announcing the same through a major news service, irrespective of whether the Company has made the required announcement; and provided, further, that no such announcement shall be made of any release or waiver granted solely to permit a transfer of securities that is not for consideration and where the transferee has agreed in writing to be bound by the terms of a Lock-Up Agreement in the form set forth as Exhibit A hereto.
The Representatives, on behalf of the several Underwriters, may, in their sole discretion, waive in writing the performance by the Company of any one or more of the foregoing covenants or extend the time for their performance.
Section 4. Payment of Expenses. The Company agrees to pay all costs, fees and expenses incurred in connection with the performance of its obligations hereunder and in connection with the transactions contemplated hereby, including without limitation (i) all expenses incident to the issuance and delivery of the Offered Shares (including all printing and engraving costs), (ii) all fees and expenses of the registrar and transfer agent of the Shares, (iii) all necessary issue, transfer and other stamp taxes in connection with the issuance and sale of the Offered Shares to the Underwriters, (iv) all fees and expenses of the Company’s counsel, independent public or certified public accountants and other advisors, (v) all costs and expenses incurred in connection with the preparation, printing, filing, shipping and distribution of the Registration Statement (including financial statements, exhibits, schedules, consents and certificates of experts), the Time of Sale Prospectus, the Prospectus, each free writing prospectus prepared by or on behalf of, used by, or referred to by the Company, and each preliminary prospectus, each Permitted Section 5(d) Communication, and all amendments and supplements thereto, and this Agreement, (vi) all filing fees, reasonable and documented attorneys’ fees and expenses incurred by the Company or the Underwriters in connection with qualifying or registering (or obtaining exemptions from the qualification
24
or registration of) all or any part of the Offered Shares for offer and sale under the state securities or blue sky laws and, if requested by the Representatives, preparing and printing a “Blue Sky Survey” or memorandum and a “Canadian wrapper”, and any supplements thereto, advising the Underwriters of such qualifications, registrations and exemptions, (vii) the costs, fees and expenses incurred by the Underwriters in connection with determining their compliance with the rules and regulations of FINRA related to the Underwriters’ participation in the offering and distribution of the Offered Shares, including any related filing fees and the legal fees of, and disbursements by, counsel to the Underwriters; provided, however, that such legal fees, taken together with the legal fees described in clause (vi) above, shall not exceed $40,000 in the aggregate, (viii) the costs and expenses of the Company relating to investor presentations on any “road show”, any Permitted Section 5(d) Communication or any Section 5(d) Oral Communication undertaken in connection with the offering of the Offered Shares, including, without limitation, expenses associated with the preparation or dissemination of any electronic road show, expenses associated with the production of road show slides and graphics, fees and expenses of any consultants engaged in connection with the road show presentations with the prior approval of the Company, travel and lodging expenses of the representatives, employees and officers of the Company and any such consultants (it being understood that the Underwriters will pay or cause to be paid the travel and lodging expenses of their representatives), and 50% of the cost of any aircraft chartered in connection with the road show, with the remaining 50% of the cost of such aircraft to be paid by the Underwriters, (ix) the fees and expenses associated with listing the Offered Shares on Nasdaq and (x) all other fees, costs and expenses of the nature referred to in Item 13 of Part II of the Registration Statement. Except as provided in this Section 4 or in Section 7, Section 9 or Section 10 hereof, the Underwriters shall pay their own expenses, including the fees and disbursements of their counsel.
Section 5. Covenant of the Underwriters. Each Underwriter severally and not jointly covenants with the Company not to take any action that would result in the Company being required to file with the Commission pursuant to Rule 433(d) under the Securities Act a free writing prospectus prepared by or on behalf of such Underwriter that otherwise would not, but for such actions, be required to be filed by the Company under Rule 433(d).
Section 6. Conditions of the Obligations of the Underwriters. The respective obligations of the several Underwriters hereunder to purchase and pay for the Offered Shares as provided herein on the First Closing Date and, with respect to the Optional Shares, each Option Closing Date, shall be subject to the accuracy of the representations and warranties on the part of the Company set forth in Section 1 hereof as of the date hereof and as of the First Closing Date as though then made and, with respect to the Optional Shares, as of each Option Closing Date as though then made, to the timely performance by the Company of its covenants and other obligations hereunder, and to each of the following additional conditions:
(a) Comfort Letter. On the date hereof, the Representatives shall have received from Ernst & Young LLP, independent registered public accountants for the Company, a letter dated the date hereof addressed to the Underwriters, in form and substance satisfactory to the Representatives, containing statements and information of the type ordinarily included in accountant’s “comfort letters” to underwriters, delivered according to Statement of Auditing Standards No. 72 (or any successor bulletin), with respect to the audited and unaudited financial statements and certain financial information contained in the Registration Statement, the Time of Sale Prospectus, and each free writing prospectus, if any.
(b) Compliance with Registration Requirements; No Stop Order; No Objection from FINRA. For the period from and after the date of this Agreement and through and including the First Closing Date and, with respect to any Optional Shares purchased after the First Closing Date, each Option Closing Date:
25
(i) The Company shall have filed the Prospectus with the Commission (including the information required by Rule 430A under the Securities Act) in the manner and within the time period required by Rule 424(b) under the Securities Act; or the Company shall have filed a post-effective amendment to the Registration Statement containing the information required by such Rule 430A, and such post-effective amendment shall have become effective.
(ii) No stop order suspending the effectiveness of the Registration Statement or any post-effective amendment to the Registration Statement shall be in effect, and no proceedings for such purpose shall have been instituted or, to the Company’s knowledge, are contemplated or threatened by the Commission.
(iii) FINRA shall have raised no objection to the fairness and reasonableness of the underwriting terms and arrangements.
(c) No Material Adverse Change. For the period from and after the date of this Agreement and through and including the First Closing Date and, with respect to any Optional Shares purchased after the First Closing Date, each Option Closing Date in the judgment of the Representatives there shall not have occurred any Material Adverse Change.
(d) Opinion and Negative Assurance Letter of Counsel for the Company. On each of the First Closing Date and each Option Closing Date, the Representatives shall have received the opinion (including tax and regulatory) and negative assurance letter of Cooley LLP, counsel for the Company, dated as of such date, in form and substance satisfactory to the Representatives.
(e) Opinion of Intellectual Property Counsel for the Company. On each of the First Closing Date and each Option Closing Date, the Representatives shall have received the opinion of Cooley LLP, counsel for the Company with respect to intellectual property matters, dated as of such date, in form and substance satisfactory to the Representatives.
(f) Opinion and Negative Assurance Letter of Counsel for the Underwriters. On each of the First Closing Date and each Option Closing Date, the Representatives shall have received the opinion and negative assurance letter of Latham & Watkins LLP, counsel for the Underwriters in connection with the offer and sale of the Offered Shares, dated as of such date, in form and substance satisfactory to the Representatives.
(g) Officers’ Certificate. On each of the First Closing Date and each Option Closing Date, the Representatives shall have received a certificate executed by the Chief Executive Officer or President of the Company and the Chief Financial Officer of the Company, dated as of such date, to the effect set forth in Section 6(b)(ii) and further to the effect that:
(i) for the period from and including the date of this Agreement through and including such date, there has not occurred any Material Adverse Change;
(ii) the representations, warranties and covenants of the Company set forth in Section 1 of this Agreement are true and correct with the same force and effect as though expressly made on and as of such date; and
(iii) the Company has complied with all the agreements hereunder and satisfied all the conditions on its part to be performed or satisfied hereunder at or prior to such date.
26
(h) Bring-down Comfort Letter. On each of the First Closing Date and each Option Closing Date, the Representatives shall have received from Ernst & Young LLP, independent registered public accountants for the Company, a letter dated such date, in form and substance satisfactory to the Representatives, which letter shall: (i) reaffirm the statements made in the letter furnished by them pursuant to Section 6(a), except that the specified date referred to therein for the carrying out of procedures shall be no more than three business days prior to the First Closing Date or the applicable Option Closing Date, as the case may be; and (ii) cover certain financial information contained in the Prospectus.
(i) Lock-Up Agreements. On or prior to the date hereof, the Company shall have furnished to the Representatives an agreement in the form of Exhibit A hereto from each director and officer of the Company and substantially all securityholders of the Company.
(j) Rule 462(b) Registration Statement. In the event that a Rule 462(b) Registration Statement is filed in connection with the offering contemplated by this Agreement, such Rule 462(b) Registration Statement shall have been filed with the Commission on the date of this Agreement and shall have become effective automatically upon such filing.
(k) Approval of Listing. At the First Closing Date, the Offered Shares shall have been approved for listing on Nasdaq, subject only to official notice of issuance.
(l) Additional Documents. On or before each of the First Closing Date and each Option Closing Date, the Representatives and counsel for the Underwriters shall have received such information, documents and opinions as they may reasonably request for the purposes of enabling them to pass upon the issuance and sale of the Offered Shares as contemplated herein, or in order to evidence the accuracy of any of the representations and warranties, or the satisfaction of any of the conditions or agreements, herein contained; and all proceedings taken by the Company in connection with the issuance and sale of the Offered Shares as contemplated herein and in connection with the other transactions contemplated by this Agreement shall be reasonably satisfactory in form and substance to the Representatives and counsel for the Underwriters.
If any condition specified in this Section 6 is not satisfied when and as required to be satisfied, and such condition has not been waived in writing by the Representatives, this Agreement may be terminated by the Representatives by notice from the Representatives to the Company at any time on or prior to the First Closing Date and, with respect to the Optional Shares, at any time on or prior to the applicable Option Closing Date, which termination shall be without liability on the part of any party to any other party, except that Section 4, Section 7, Section 9 and Section 10 shall at all times be effective and shall survive such termination.
Section 7. Reimbursement of Underwriters’ Expenses. If this Agreement is terminated by the Representatives pursuant to Section 6, Section 11 or Section 12, or if the sale to the Underwriters of the Offered Shares on the First Closing Date is not consummated because of any refusal, inability or failure on the part of the Company to perform any agreement herein or to comply with any provision hereof, the Company agrees to reimburse the Representatives and the other Underwriters (or such Underwriters as have terminated this Agreement with respect to themselves), severally, upon demand for all documented out-of-pocket expenses that shall have been reasonably incurred by the Representatives and the Underwriters in connection with the proposed purchase and the offering and sale of the Offered Shares, including, but not limited to, reasonable and documented fees and disbursements of counsel, printing expenses, travel expenses, postage, facsimile and telephone charges. For the avoidance of doubt, it is understood that the Company will not pay or reimburse any costs, fees or expenses solely attributable to any Underwriter that defaults on its obligations to purchase the Offered Shares pursuant to Section 11; provided that the foregoing shall not relieve the Company of its obligation to reimburse any costs, fees or expenses that constitute general syndicate expenses, notwithstanding that such expenses may have been initially incurred by or through such defaulting Underwriter.
27
Section 8. Effectiveness of this Agreement. This Agreement shall become effective upon the execution and delivery hereof by the parties hereto.
Section 9. Indemnification.
(a) Indemnification of the Underwriters. The Company agrees to indemnify and hold harmless each Underwriter, its affiliates, directors, officers, employees and agents, and each person, if any, who controls any Underwriter within the meaning of the Securities Act or the Exchange Act against any loss, claim, damage, liability or expense, as incurred, to which such Underwriter or such affiliate, director, officer, employee, agent or controlling person may become subject, under the Securities Act, the Exchange Act, other federal or state statutory law or regulation, or the laws or regulations of foreign jurisdictions where Offered Shares have been offered or sold or at common law or otherwise (including in settlement of any litigation, if such settlement is effected with the written consent of the Company), insofar as such loss, claim, damage, liability or expense (or actions in respect thereof as contemplated below) arises out of or is based upon (A) (i) any untrue statement or alleged untrue statement of a material fact contained in the Registration Statement, or any amendment thereto, or the omission or alleged omission to state therein a material fact required to be stated therein or necessary to make the statements therein not misleading; or (ii) any untrue statement or alleged untrue statement of a material fact included in any preliminary prospectus, the Time of Sale Prospectus, any free writing prospectus that the Company has used, referred to or filed, or is required to file, pursuant to Rule 433(d) of the Securities Act, any Marketing Material, any Section 5(d) Written Communication or the Prospectus (or any amendment or supplement to the foregoing), or the omission or alleged omission to state therein a material fact necessary in order to make the statements, in the light of the circumstances under which they were made, not misleading; or (iii) any act or failure to act or any alleged act or failure to act by any Underwriter in connection with, or relating in any manner to, the Shares or the offering contemplated hereby, and which is included as part of or referred to in any loss, claim, damage, liability or action arising out of or based upon any matter covered by clause (i) or (ii) above, or (B) the violation of any laws or regulations of foreign jurisdictions where Offered Shares have been offered or sold; and to reimburse each Underwriter and each such affiliate, director, officer, employee, agent and controlling person for any and all reasonable expenses (including the fees and disbursements of counsel) as such expenses are incurred by such Underwriter or such affiliate, director, officer, employee, agent or controlling person in connection with investigating, defending, settling, compromising or paying any such loss, claim, damage, liability, expense or action; provided, however, that the foregoing indemnity agreement shall not apply to any loss, claim, damage, liability or expense to the extent, but only to the extent, arising out of or based upon any untrue statement or alleged untrue statement or omission or alleged omission made in reliance upon and in conformity with information relating to any Underwriter furnished to the Company by the Representatives in writing expressly for use in the Registration Statement, any preliminary prospectus, the Time of Sale Prospectus, any such free writing prospectus, any Marketing Material, any Section 5(d) Written Communication or the Prospectus (or any amendment or supplement thereto), it being understood and agreed that the only such information consists of the information described in Section 9(b) below. The indemnity agreement set forth in this Section 9(a) shall be in addition to any liabilities that the Company may otherwise have.
(b) Indemnification of the Company, its Directors and Officers. Each Underwriter agrees, severally and not jointly, to indemnify and hold harmless the Company, each of its directors, each of its officers who signed the Registration Statement and each person, if any, who controls the Company within the meaning of the Securities Act or the Exchange Act, against any loss, claim, damage, liability or expense, as incurred, to which the Company, or any such director, officer or controlling person may become subject, under the Securities Act, the Exchange Act, or other federal or state statutory law or
28
regulation, or at common law or otherwise (including in settlement of any litigation, if such settlement is effected with the written consent of such Underwriter), insofar as such loss, claim, damage, liability or expense (or actions in respect thereof as contemplated below) arises out of or is based upon (i) any untrue statement or alleged untrue statement of a material fact contained in the Registration Statement, or any amendment thereto, or any omission or alleged omission to state therein a material fact required to be stated therein or necessary to make the statements therein not misleading or (ii) any untrue statement or alleged untrue statement of a material fact included in any preliminary prospectus, the Time of Sale Prospectus, any free writing prospectus, that the Company has used, referred to or filed, or is required to file, pursuant to Rule 433 of the Securities Act, any Section 5(d) Written Communication or the Prospectus (or any such amendment or supplement) or the omission or alleged omission to state therein a material fact necessary in order to make the statements, in the light of the circumstances under which they were made, not misleading, in each case to the extent, but only to the extent, that such untrue statement or alleged untrue statement or omission or alleged omission was made in the Registration Statement, such preliminary prospectus, the Time of Sale Prospectus, such free writing prospectus, such Section 5(d) Written Communication or the Prospectus (or any such amendment or supplement), in reliance upon and in conformity with information relating to such Underwriter furnished to the Company by the Representatives in writing expressly for use therein; and to reimburse the Company, or any such director, officer or controlling person for any and all reasonable expenses (including the fees and disbursements of counsel) as such expenses are incurred by the Company, or any such director, officer or controlling person in connection with investigating, defending, settling, compromising or paying any such loss, claim, damage, liability, expense or action. The Company hereby acknowledges that the only information that the Representatives have furnished to the Company expressly for use in the Registration Statement, any preliminary prospectus, the Time of Sale Prospectus, any free writing prospectus that the Company has filed, or is required to file, pursuant to Rule 433(d) of the Securities Act, any Section 5(d) Written Communication or the Prospectus (or any amendment or supplement to the foregoing) are the statements set forth in the first sentence of the third paragraph under the caption “Underwriting”, the first sentence of the first paragraph under the caption “Commission and Expenses”, and the first sentence of the first paragraph under the caption “Stabilization”, in each case in the section titled “Underwriting” in the Preliminary Prospectus and the Prospectus. The indemnity agreement set forth in this Section 9(b) shall be in addition to any liabilities that each Underwriter may otherwise have.
(c) Notifications and Other Indemnification Procedures. Promptly after receipt by an indemnified party under this Section 9 of notice of the commencement of any action, such indemnified party will, if a claim in respect thereof is to be made against an indemnifying party under this Section 9, notify the indemnifying party in writing of the commencement thereof, but the omission to so notify the indemnifying party will not relieve the indemnifying party from any liability which it may have to any indemnified party to the extent the indemnifying party is not materially prejudiced as a proximate result of such failure and shall not in any event relieve the indemnifying party from any liability that it may have otherwise than on account of this indemnity agreement. In case any such action is brought against any indemnified party and such indemnified party seeks or intends to seek indemnity from an indemnifying party, the indemnifying party will be entitled to participate in, and, to the extent that it shall elect, jointly with all other indemnifying parties similarly notified, by written notice delivered to the indemnified party promptly after receiving the aforesaid notice from such indemnified party, to assume the defense thereof with counsel reasonably satisfactory to such indemnified party; provided, however, that if the defendants in any such action include both the indemnified party and the indemnifying party and the indemnified party shall have reasonably concluded that a conflict may arise between the positions of the indemnifying party and the indemnified party in conducting the defense of any such action or that there may be legal defenses available to it and/or other indemnified parties which are different from or additional to those available to the indemnifying party, the indemnified party or parties shall have the right to select separate counsel to assume such legal defenses and to otherwise participate in the defense of such action on behalf of such indemnified party or parties. Upon receipt of notice from the indemnifying party to such indemnified
29
party of such indemnifying party’s election to so assume the defense of such action and approval by the indemnified party of counsel, the indemnifying party will not be liable to such indemnified party under this Section 9 for any reasonable and documented legal or other expenses subsequently incurred by such indemnified party in connection with the defense thereof unless (i) the indemnified party shall have employed separate counsel in accordance with the proviso to the preceding sentence (it being understood, however, that the indemnifying party shall not be liable for the fees and expenses of more than one separate counsel (together with local counsel), representing the indemnified parties who are parties to such action), which counsel (together with any local counsel) for the indemnified parties shall be selected by the Representatives (in the case of counsel for the indemnified parties referred to in Section 9(a) above) or by the Company (in the case of counsel for the indemnified parties referred to in Section 9(b) above)) or (ii) the indemnifying party shall not have employed counsel reasonably satisfactory to the indemnified party to represent the indemnified party within a reasonable time after notice of commencement of the action or (iii) the indemnifying party has authorized in writing the employment of counsel for the indemnified party at the expense of the indemnifying party, in each of which cases the fees and expenses of counsel shall be at the expense of the indemnifying party and shall be paid as they are incurred.
(d) Settlements. The indemnifying party under this Section 9 shall not be liable for any settlement of any proceeding effected without its written consent, but if settled with such consent or if there be a final judgment for the plaintiff, the indemnifying party agrees to indemnify the indemnified party against any loss, claim, damage, liability or expense by reason of such settlement or judgment. Notwithstanding the foregoing sentence, if at any time an indemnified party shall have requested an indemnifying party to reimburse the indemnified party for the reasonable and documented fees and expenses of counsel as contemplated by Section 9(c) hereof, the indemnifying party shall be liable for any settlement of any proceeding effected without its written consent if (i) such settlement is entered into more than 30 days after receipt by such indemnifying party of the aforesaid request and (ii) such indemnifying party shall not have reimbursed the indemnified party in accordance with such request prior to the date of such settlement. No indemnifying party shall, without the prior written consent of the indemnified party, effect any settlement, compromise or consent to the entry of judgment in any pending or threatened action, suit or proceeding in respect of which any indemnified party is or could have been a party and indemnity was or could have been sought hereunder by such indemnified party, unless such settlement, compromise or consent includes an unconditional release of such indemnified party from all liability on claims that are the subject matter of such action, suit or proceeding and does not include an admission of fault or culpability or a failure to act by or on behalf of such indemnified party.
Section 10. Contribution. If the indemnification provided for in Section 9 is for any reason held to be unavailable to or otherwise insufficient to hold harmless an indemnified party in respect of any losses, claims, damages, liabilities or expenses referred to therein, then each indemnifying party shall contribute to the aggregate amount paid or payable by such indemnified party, as incurred, as a result of any losses, claims, damages, liabilities or expenses referred to therein (i) in such proportion as is appropriate to reflect the relative benefits received by the Company, on the one hand, and the Underwriters, on the other hand, from the offering of the Offered Shares pursuant to this Agreement or (ii) if the allocation provided by clause (i) above is not permitted by applicable law, in such proportion as is appropriate to reflect not only the relative benefits referred to in clause (i) above but also the relative fault of the Company, on the one hand, and the Underwriters, on the other hand, in connection with the statements or omissions which resulted in such losses, claims, damages, liabilities or expenses, as well as any other relevant equitable considerations. The relative benefits received by the Company on the one hand, and the Underwriters, on the other hand, in connection with the offering of the Offered Shares pursuant to this Agreement shall be deemed to be in the same respective proportions as the total proceeds from the offering of the Offered Shares pursuant to this Agreement (before deducting expenses) received by the Company, and the total underwriting discounts and commissions received by the Underwriters, in each case as set forth on the front cover page of the Prospectus, bear to the aggregate initial public offering
30
price of the Offered Shares as set forth on such cover. The relative fault of the Company, on the one hand, and the Underwriters, on the other hand, shall be determined by reference to, among other things, whether any such untrue or alleged untrue statement of a material fact or omission or alleged omission to state a material fact relates to information supplied by the Company, on the one hand, or the Underwriters, on the other hand, and the parties’ relative intent, knowledge, access to information and opportunity to correct or prevent such statement or omission.
The amount paid or payable by a party as a result of the losses, claims, damages, liabilities and expenses referred to above shall be deemed to include, subject to the limitations set forth in Section 9(c), any legal or other fees or expenses reasonably incurred by such party in connection with investigating or defending any action or claim. The provisions set forth in Section 9(c) with respect to notice of commencement of any action shall apply if a claim for contribution is to be made under this Section 10; provided, however, that no additional notice shall be required with respect to any action for which notice has been given under Section 9(c) for purposes of indemnification.
The Company and the Underwriters agree that it would not be just and equitable if contribution pursuant to this Section 10 were determined by pro rata allocation (even if the Underwriters were treated as one entity for such purpose) or by any other method of allocation which does not take account of the equitable considerations referred to in this Section 10.
Notwithstanding the provisions of this Section 10, no Underwriter shall be required to contribute any amount in excess of the underwriting discounts and commissions received by such Underwriter in connection with the Offered Shares underwritten by it and distributed to the public. No person guilty of fraudulent misrepresentation (within the meaning of Section 11(f) of the Securities Act) shall be entitled to contribution from any person who was not guilty of such fraudulent misrepresentation. The Underwriters’ obligations to contribute pursuant to this Section 10 are several, and not joint, in proportion to their respective underwriting commitments as set forth opposite their respective names on Schedule A. For purposes of this Section 10, each affiliate, director, officer, employee and agent of an Underwriter and each person, if any, who controls an Underwriter within the meaning of the Securities Act or the Exchange Act shall have the same rights to contribution as such Underwriter, and each director of the Company, each officer of the Company who signed the Registration Statement, and each person, if any, who controls the Company within the meaning of the Securities Act and the Exchange Act shall have the same rights to contribution as the Company.
Section 11. Default of One or More of the Several Underwriters. If, on the First Closing Date or any Option Closing Date any one or more of the several Underwriters shall fail or refuse to purchase Offered Shares that it or they have agreed to purchase hereunder on such date, and the aggregate number of Offered Shares which such defaulting Underwriter or Underwriters agreed but failed or refused to purchase does not exceed 10% of the aggregate number of the Offered Shares to be purchased on such date, the Representatives may make arrangements satisfactory to the Company for the purchase of such Offered Shares by other persons, including any of the Underwriters, but if no such arrangements are made by such date, the other Underwriters shall be obligated, severally and not jointly, in the proportions that the number of Firm Shares set forth opposite their respective names on Schedule A bears to the aggregate number of Firm Shares set forth opposite the names of all such non-defaulting Underwriters, or in such other proportions as may be specified by the Representatives with the consent of the non-defaulting Underwriters, to purchase the Offered Shares which such defaulting Underwriter or Underwriters agreed but failed or refused to purchase on such date. If, on the First Closing Date or any Option Closing Date any one or more of the Underwriters shall fail or refuse to purchase Offered Shares and the aggregate number of Offered Shares with respect to which such default occurs exceeds 10% of the aggregate number of Offered Shares to be purchased on such date, and arrangements satisfactory to the Representatives and the Company for the purchase of such Offered Shares are not made within 48 hours after such default, this
31
Agreement shall terminate without liability of any party (other than the breaching Underwriter or Underwriters) to any other party except that the provisions of Section 4, Section 7, Section 9 and Section 10 shall at all times be effective and shall survive such termination. In any such case either the Representatives or the Company shall have the right to postpone the First Closing Date or the applicable Option Closing Date, as the case may be, but in no event for longer than seven days in order that the required changes, if any, to the Registration Statement and the Prospectus or any other documents or arrangements may be effected.
As used in this Agreement, the term “Underwriter” shall be deemed to include any person substituted for a defaulting Underwriter under this Section 11. Any action taken under this Section 11 shall not relieve any defaulting Underwriter from liability in respect of any default of such Underwriter under this Agreement.
Section 12. Termination of this Agreement. Prior to the purchase of the Firm Shares by the Underwriters on the First Closing Date, this Agreement may be terminated by the Representatives by notice given to the Company if at any time: (i) trading or quotation in any of the Company’s securities shall have been suspended or limited by the Commission or by Nasdaq, or trading in securities generally on either Nasdaq or the New York Stock Exchange shall have been suspended or limited, or minimum or maximum prices shall have been generally established on any of such stock exchanges; (ii) a general banking moratorium shall have been declared by any federal, New York or California authorities; (iii) there shall have occurred any outbreak or escalation of national or international hostilities or any crisis or calamity, or any change in the United States or international financial markets, or any substantial change or development involving a prospective substantial change in United States’ or international political, financial or economic conditions, as in the judgment of the Representatives is material and adverse and makes it impracticable to market the Offered Shares in the manner and on the terms described in the Time of Sale Prospectus or the Prospectus or to enforce contracts for the sale of securities; (iv) in the judgment of the Representatives there shall have occurred any Material Adverse Change; or (v) the Company shall have sustained a loss by strike, fire, flood, earthquake, accident or other calamity of such character as in the judgment of the Representatives may interfere materially with the conduct of the business and operations of the Company regardless of whether or not such loss shall have been insured. Any termination pursuant to this Section 12 shall be without liability on the part of (a) the Company to any Underwriter, except that the Company shall be obligated to reimburse the expenses of the Representatives and the Underwriters pursuant to Section 4 or Section 7 hereof or (b) any Underwriter to the Company; provided, however, that the provisions of Section 9 and Section 10 shall at all times be effective and shall survive such termination.
Section 13. No Advisory or Fiduciary Relationship. The Company acknowledges and agrees that (a) the purchase and sale of the Offered Shares pursuant to this Agreement, including the determination of the public offering price of the Offered Shares and any related discounts and commissions, is an arm’s-length commercial transaction between the Company, on the one hand, and the several Underwriters, on the other hand, and does not constitute a recommendation, investment advice, or solicitation of any action by the Underwriters, (b) in connection with the offering contemplated hereby and the process leading to such transaction, each Underwriter is and has been acting solely as a principal and is not the agent or fiduciary of the Company, or its stockholders, creditors, employees or any other party, (c) no Underwriter has assumed or will assume an advisory or fiduciary responsibility in favor of the Company with respect to the offering contemplated hereby or the process leading thereto (irrespective of whether such Underwriter has advised or is currently advising the Company on other matters) and no Underwriter has any obligation to the Company with respect to the offering contemplated hereby except the obligations expressly set forth in this Agreement, (d) the Underwriters and their respective affiliates may be engaged in a broad range of transactions that involve interests that differ from those of the Company, (e) the Underwriters have not provided any legal, accounting, regulatory or tax advice with
32
respect to the offering contemplated hereby and the Company has consulted its own legal, accounting, regulatory and tax advisors to the extent it deemed appropriate and (f) none of the activities of the Underwriters in connection with the transaction contemplated herein constitutes a recommendation, investment advice, or solicitation of any action by the Underwriters with respect to any entity or natural person.
Section 14. Representations and Indemnities to Survive Delivery. The respective indemnities, agreements, representations, warranties and other statements of the Company, of its officers and of the several Underwriters set forth in or made pursuant to this Agreement will remain in full force and effect, regardless of any investigation made by or on behalf of any Underwriter or the Company or any of its or their partners, officers or directors or any controlling person, as the case may be, and, anything herein to the contrary notwithstanding, will survive delivery of and payment for the Offered Shares sold hereunder and any termination of this Agreement.
Section 15. Notices. All communications hereunder shall be in writing and shall be mailed, hand delivered or telecopied and confirmed to the parties hereto as follows:
| If to the Representatives: | Jefferies LLC | |
| 520 Madison Avenue | ||
| New York, New York 10022 | ||
| Attention: General Counsel | ||
| TD Securities (USA) LLC | ||
| 1 Vanderbilt Avenue | ||
| New York, New York 10017 | ||
| Attention: Head of Equity Capital Markets; General Counsel | ||
| Email: ciblegal@tdsecurities.com | ||
| Evercore Group L.L.C. | ||
| 55 East 52nd Street, 35th Floor | ||
| New York, New York 10055 | ||
| Attention: ECM General Counsel | ||
| Cantor Fitzgerald & Co. | ||
| 110 East 59th Street, 6th Floor | ||
| New York, New York 10022 | ||
| Attention: General Counsel | ||
| with a copy to: | Latham & Watkins LLP | |
| 12670 High Bluff Drive | ||
| San Diego, CA 92130 | ||
| Attention: Matthew T. Bush | ||
| Electra Therapeutics, Inc. | ||
| 230 E Grand Avenue, Suite S-100 | ||
| South San Francisco, California 94080 | ||
| Attention: Chief Executive Officer | ||
| with copies to: | Electra Therapeutics, Inc. | |
| legalnotices@electra-therapeutics.com | ||
33
| Cooley LLP 10265 Science Center Drive | ||
| San Diego, California 92121 | ||
| Facsimile: (858) 550-6420 | ||
| Attention: Charles Bair, Charles S. Kim and Sara Semnani |
Any party hereto may change the address for receipt of communications by giving written notice to the others.
Section 16. Successors. This Agreement will inure to the benefit of and be binding upon the parties hereto, including any substitute Underwriters pursuant to Section 11 hereof, and to the benefit of the affiliates, directors, officers, employees, agents and controlling persons referred to in Section 9 and Section 10, and in each case their respective successors, and personal representatives, and no other person will have any right or obligation hereunder. The term “successors” shall not include any purchaser of the Offered Shares as such from any of the Underwriters merely by reason of such purchase.
Section 17. Partial Unenforceability. The invalidity or unenforceability of any section, paragraph or provision of this Agreement shall not affect the validity or enforceability of any other section, paragraph or provision hereof. If any section, paragraph or provision of this Agreement is for any reason determined to be invalid or unenforceable, there shall be deemed to be made such minor changes (and only such minor changes) as are necessary to make it valid and enforceable.
Section 18. Recognition of the U.S. Special Resolution Regimes.
(a) In the event that any Underwriter that is a Covered Entity becomes subject to a proceeding under a U.S. Special Resolution Regime, the transfer from such Underwriter of this Agreement, and any interest and obligation in or under this Agreement, will be effective to the same extent as the transfer would be effective under the U.S. Special Resolution Regime if this Agreement, and any such interest and obligation, were governed by the laws of the United States or a state of the United States.
(b) In the event that any Underwriter that is a Covered Entity or a BHC Act Affiliate of such Underwriter becomes subject to a proceeding under a U.S. Special Resolution Regime, Default Rights under this Agreement that may be exercised against such Underwriter are permitted to be exercised to no greater extent than such Default Rights could be exercised under the U.S. Special Resolution Regime if this Agreement were governed by the laws of the United States or a state of the United States.
For purposes of this Agreement, (A) “BHC Act Affiliate” has the meaning assigned to the term “affiliate” in, and shall be interpreted in accordance with, 12 U.S.C. § 1841(k); (B) “Covered Entity” means any of the following: (i) a “covered entity” as that term is defined in, and interpreted in accordance with, 12 C.F.R. § 252.82(b); (ii) a “covered bank” as that term is defined in, and interpreted in accordance with, 12 C.F.R. § 47.3(b); or (iii) a “covered FSI” as that term is defined in, and interpreted in accordance with, 12 C.F.R. § 382.2(b); (C) “Default Right” has the meaning assigned to that term in, and shall be interpreted in accordance with, 12 C.F.R. §§ 252.81, 47.2 or 382.1, as applicable; and (D) “U.S. Special Resolution Regime” means each of (i) the Federal Deposit Insurance Act and the regulations promulgated thereunder and (ii) Title II of the Dodd-Frank Wall Street Reform and Consumer Protection Act and the regulations promulgated thereunder.
34
Section 19. Governing Law Provisions. This Agreement shall be governed by and construed in accordance with the internal laws of the State of New York applicable to agreements made and to be performed in such state. Any legal suit, action or proceeding arising out of or based upon this Agreement or the transactions contemplated hereby may be instituted in the federal courts of the United States of America located in the Borough of Manhattan in the City of New York or the courts of the State of New York in each case located in the Borough of Manhattan in the City of New York (collectively, the “Specified Courts”), and each party irrevocably submits to the exclusive jurisdiction (except for proceedings instituted in regard to the enforcement of a judgment of any such court, as to which such jurisdiction is non-exclusive) of such courts in any such suit, action or proceeding. Service of any process, summons, notice or document by mail to such party’s address set forth above shall be effective service of process for any suit, action or other proceeding brought in any such court. The parties irrevocably and unconditionally waive any objection to the laying of venue of any suit, action or other proceeding in the Specified Courts and irrevocably and unconditionally waive and agree not to plead or claim in any such court that any such suit, action or other proceeding brought in any such court has been brought in an inconvenient forum.
Section 20. General Provisions. This Agreement constitutes the entire agreement of the parties to this Agreement and supersedes all prior written or oral and all contemporaneous oral agreements, understandings and negotiations with respect to the subject matter hereof. This Agreement may be executed in two or more counterparts, each one of which shall be an original, with the same effect as if the signatures thereto and hereto were upon the same instrument. In the event that any signature is delivered by facsimile transmission or by e-mail delivery of a “.pdf” format data file, such signature shall create a valid and binding obligation of the party executing (or on whose behalf such signature is executed) with the same force and effect as if such facsimile or “.pdf” signature page were an original thereof. Counterparts may be delivered via facsimile, electronic mail (including any electronic signature covered by the U.S. federal ESIGN Act of 2000, Uniform Electronic Transactions Act, the Electronic Signatures and Records Act or other applicable law, e.g., www.docusign.com) or other transmission method and any counterpart so delivered shall be deemed to have been duly and validly delivered and be valid and effective for all purposes. This Agreement may not be amended or modified unless in writing by all of the parties hereto, and no condition herein (express or implied) may be waived unless waived in writing by each party whom the condition is meant to benefit. The section headings herein are for the convenience of the parties only and shall not affect the construction or interpretation of this Agreement.
Each of the parties hereto acknowledges that it is a sophisticated business person who was adequately represented by counsel during negotiations regarding the provisions hereof, including, without limitation, the indemnification provisions of Section 9 and the contribution provisions of Section 10, and is fully informed regarding said provisions. Each of the parties hereto further acknowledges that the provisions of Section 9 and Section 10 hereof fairly allocate the risks in light of the ability of the parties to investigate the Company, its affairs and its business in order to assure that adequate disclosure has been made in the Registration Statement, any preliminary prospectus, the Time of Sale Prospectus, each free writing prospectus and the Prospectus (and any amendments and supplements to the foregoing), as contemplated by the Securities Act and the Exchange Act.
35
Exhibit 1.1
If the foregoing is in accordance with your understanding of our agreement, kindly sign and return to the Company the enclosed copies hereof, whereupon this instrument, along with all counterparts hereof, shall become a binding agreement in accordance with its terms.
| Very truly yours, | ||
| ELECTRA THERAPEUTICS, INC. | ||
| By: |
| |
| Name: | ||
| Title: | ||
The foregoing Underwriting Agreement is hereby confirmed and accepted by the Representatives in New York, New York as of the date first above written.
| JEFFERIES LLC | ||
| TD SECURITIES (USA) LLC | ||
| EVERCORE GROUP L.L.C. | ||
| CANTOR FITZGERALD & CO. | ||
| Acting individually and as Representatives of the several Underwriters named in the attached Schedule A. | ||
| JEFFERIES LLC | ||
| By: |
| |
| Name: | ||
| Title: | ||
| TD SECURITIES (USA) LLC | ||
| By: |
| |
| Name: | ||
| Title: | ||
| EVERCORE GROUP L.L.C. | ||
| By: |
| |
| Name: | ||
| Title: | ||
| CANTOR FITZGERALD & CO. | ||
| By: |
| |
| Name: | ||
| Title: | ||
Schedule A
| Underwriters | Number of Firm Shares to be Purchased |
|||
| Jefferies LLC |
[ | •] | ||
| TD Securities (USA) LLC |
[ | •] | ||
| Evercore Group L.L.C. |
[ | •] | ||
| Cantor Fitzgerald & Co. |
[ | •] | ||
|
|
|
|||
| Total |
[ | •] | ||
|
|
|
|||
Schedule B
Free Writing Prospectuses Included in the Time of Sale Prospectus
[None.]
Schedule C
Permitted Section 5(d) Communications
All written communications presented to potential investors in reliance on Section 5(d) of the U.S. Securities Act of 1933, as amended, and submitted to the U.S. Securities and Exchange Commission.
Schedule D
Pricing Terms
Firm Shares –
Optional Shares –
Price per Share to Public –
D-1
Exhibit A
Form of Lock-up Agreement
___________, 2026
Jefferies LLC
TD Securities (USA) LLC
Evercore Group L.L.C.
Cantor Fitzgerald & Co.
As Representatives of the Several Underwriters
c/o Jefferies LLC
520 Madison Avenue
New York, New York 10022
c/o TD Securities (USA) LLC
1 Vanderbilt Avenue
New York, New York 10017
c/o Evercore Group L.L.C.
55 E. 52nd St.
New York, New York 10055
c/o Cantor Fitzgerald & Co.
110 E. 59th St., 6th Floor
New York, New York 10020
RE: Electra Therapeutics, Inc. (the “Company”)
Ladies & Gentlemen:
The undersigned is an owner of shares of common stock, par value $0.0001 per share, of the Company (“Shares”) or of securities convertible into or exchangeable or exercisable for Shares. The Company proposes to conduct a public offering of Shares (the “Offering”) for which Jefferies LLC (“Jefferies”), TD Securities (USA) LLC (“TD Cowen”), Evercore Group L.L.C. and Cantor Fitzgerald & Co. will act as the representatives (collectively, the “Representatives”) of the underwriters. The undersigned recognizes that the Offering will benefit each of the Company and the undersigned. The undersigned acknowledges that the underwriters are relying on the representations and agreements of the undersigned contained in this agreement in conducting the Offering and, at a subsequent date, in entering into an underwriting agreement (the “Underwriting Agreement”) and other underwriting arrangements with the Company with respect to the Offering.
Annex A sets forth definitions for capitalized terms used in this agreement that are not defined in the body of this agreement. Those definitions are a part of this agreement.
A -1
In consideration of the foregoing, and for other good and valuable consideration, the receipt and sufficiency of which are hereby acknowledged, the undersigned hereby agrees that, during the Lock-up Period, the undersigned will not (and will cause any Family Member not to), subject to the exceptions set forth in this agreement, without the prior written consent of Jefferies and TD Cowen (together, the “Lock-up Representatives”), which may withhold their consent in their sole discretion:
| • | Sell or Offer to Sell any Shares or Related Securities currently or hereafter owned either of record or beneficially (as defined in Rule 13d-3 under the Exchange Act) by the undersigned or such Family Member; |
| • | enter into any Swap; |
| • | make any demand for, or exercise any right with respect to, the registration under the Securities Act of the offer and sale of any Shares or Related Securities, or cause to be filed a registration statement, prospectus or prospectus supplement (or an amendment or supplement thereto) with respect to any such registration; or |
| • | publicly announce any intention to do any of the foregoing. |
The foregoing restrictions will not apply to the registration of the offer and sale of the Shares, and the sale of the Shares to the underwriters, in each case as contemplated by the Underwriting Agreement. In addition, the foregoing restrictions shall not apply to the transfer of Shares or Related Securities:
| (i) | by a bona fide gift or gifts or charitable contributions, or for bona fide estate planning purposes; |
| (ii) | by will or intestate succession upon the death of the undersigned; |
| (iii) | to any trust for the direct or indirect benefit of the undersigned or Family Member; |
| (iv) | to any Family Member; |
| (v) | by operation of law pursuant to a court order or a settlement agreement related to the distribution of assets in connection with the dissolution of a marriage or civil union; |
| (vi) | to any corporation, partnership, limited liability company or other entity all of the beneficial ownership interests of which, in each case, are held by the undersigned; |
| (vii) | in connection with a distribution or other transfer by a partnership to its partners or former partners or by a limited liability company to its members (including the contemplated distribution by Electra Therapeutics LLC of shares of Series A Preferred Stock of the Company to its equityholders prior to the closing of the Offering) or retired members or by a corporation to its stockholders or former stockholders or to any wholly owned subsidiary of such corporation; |
| (viii) | to any Affiliate of the undersigned, including investment funds or other entities under common control or management that are Affiliates of the undersigned; |
| (ix) | if the undersigned is not an officer or director of the Company, transactions relating to Shares or Related Securities acquired in the Offering or in open market transactions on or after the completion of the Offering; |
A-2
| (x) | to the Company in connection with the exercise, vesting, exchange or settlement of options, warrants or other rights to acquire Shares, including any security convertible into, exchangeable for or that represent the right to receive Shares, in accordance with their terms (including the vesting or settlement of restricted stock units and including, in each case, by way of net exercise and/or to cover withholding tax obligations in connection with such exercise, vesting, exchange or settlement) pursuant to an employee benefit plan, option, warrant or other right disclosed in the Prospectus for the Offering; |
| (xi) | to the Company in connection with (A) the termination of the undersigned’s employment with the Company or (B) any agreements under which the Company has the option to repurchase such shares; |
| (xii) | upon exercise by the undersigned of any option to purchase any Shares or the settlement of restricted stock units or other equity awards pursuant to any stock incentive plan or stock purchase plan of the Company disclosed in the Prospectus for the Offering, provided that the underlying Shares shall continue to be subject to the restrictions on transfer set forth in this agreement; |
| (xiii) | in connection with the conversion of the outstanding preferred stock of the Company into Shares, provided that any such Shares received upon such conversion shall be subject to the restrictions on transfer set forth in this agreement; and |
| (xiv) | pursuant to a bona fide third-party tender offer for securities of the Company, merger, consolidation or other similar transaction made to all holders of the Company’s securities involving a Change of Control, which transaction is approved by the Board of Directors of the Company, provided that it shall be a condition of the transfer that if the tender offer, merger, consolidation or other such transaction is not completed, the undersigned’s securities, subject to this agreement, shall remain subject to the restrictions set forth herein; |
provided, however, that in any such case, it shall be a condition to such transfer that:
| • | in the case of any transfer pursuant to clauses (i) through (viii) above, each transferee executes and delivers to the Representatives an agreement in form and substance satisfactory to the Lock-up Representatives stating that such transferee is receiving and holding such Shares and/or Related Securities subject to the provisions of this agreement and agrees not to, during the Lock-up Period, Sell or Offer to Sell such Shares and/or Related Securities, engage in any Swap or engage in any other activities restricted under this agreement except in accordance with this agreement (as if such transferee had been an original signatory hereto), and such transfer shall not involve a disposition for value; |
| • | in the case of any transfer pursuant to clauses (vi) through (ix) above, prior to the expiration of the Lock-up Period, no public disclosure or filing under the Exchange Act by any party to the transfer (donor, donee, transferor or transferee) shall be required or made voluntarily during the Lock-up Period reporting a reduction in beneficial ownership of Shares or Related Securities; and |
| • | in the case of any transfer pursuant to clauses (i) through (v) and (x) through (xiii) above, prior to the expiration of the Lock-up Period, no public disclosure or filing under the Exchange Act by any party to the transfer (donor, donee, transferor or transferee) shall be made voluntarily during the Lock-up Period, and if the undersigned is required to file a report under Section 16 of the Exchange Act or any other public filing, report or announcement legally required during the Lock-up Period reporting a change in beneficial ownership of Shares or Related Securities during the Lock-up Period, the undersigned shall include a statement in such report indicating the circumstances of such transfer. |
A-3
If the undersigned is an officer or director of the Company, the undersigned further agrees that the foregoing provisions shall be equally applicable to any Company-directed Shares the undersigned may purchase or otherwise receive in the Offering (including pursuant to a directed share program).
Furthermore, notwithstanding the restrictions imposed by this agreement, the undersigned may establish or amend a written trading plan meeting the requirements of Rule 10b5-1 under the Exchange Act relating to the transfer, sale or other disposition of Shares or Related Securities, if then permitted by the Company, provided that such plan does not provide for the sale of Shares or Related Securities during the Lock-up Period and any required public disclosure, announcement or filing under the Exchange Act made by the Company or any person regarding the establishment or amendment of such plan during the Lock-up Period shall include a statement that the undersigned is not permitted to sell securities under such plan during the Lock-up Period in contravention of this agreement, and no public announcement, report or filing under the Exchange Act, or any other public filing, report or announcement, shall be voluntarily made regarding the establishment or amendment of such plan during the Lock-up Period.
In addition, if the undersigned is an officer or director of the Company, (i) the Lock-up Representatives agree that, at least three business days before the effective date of any release or waiver of the foregoing restrictions in connection with a transfer of Shares, the Lock-up Representatives will notify the Company of the impending release or waiver, and (ii) the Company (in accordance with the provisions of the Underwriting Agreement) will announce the impending release or waiver by press release through a major news service at least two business days before the effective date of the release or waiver. Any release or waiver granted by the Lock-up Representatives hereunder to any such officer or director shall only be effective two business days after the publication date of such press release. The provisions of this paragraph will not apply if both (a) the release or waiver is effected solely to permit a transfer not for consideration and (b) the transferee has agreed in writing to be bound by the same terms described in this agreement that are applicable to the transferor to the extent and for the duration that such terms remain in effect at the time of the transfer.
The undersigned also agrees and consents to the entry of stop transfer instructions with the Company’s transfer agent and registrar against the transfer of Shares or Related Securities held by the undersigned and the undersigned’s Family Members, if any, except in compliance with the foregoing restrictions.
With respect to the Offering only, the undersigned waives any registration rights relating to registration under the Securities Act of the offer and sale of any Shares and/or any Related Securities owned either of record or beneficially by the undersigned, including any rights to receive notice of the Offering.
The undersigned confirms that the undersigned has not, and has no knowledge that any Family Member has, directly or indirectly, taken any action designed to or that might reasonably be expected to cause or result in the stabilization or manipulation of the price of any security of the Company to facilitate the sale of the Shares. The undersigned will not, and will use reasonable best efforts to cause any Family Member not to take, directly or indirectly, any such action.
The undersigned acknowledges and agrees that the underwriters have not provided any recommendation or investment advice nor have the underwriters solicited any action from the undersigned with respect to the Offering and the undersigned has consulted their own legal, accounting, financial, regulatory and tax advisors to the extent deemed appropriate. The undersigned further acknowledges and agrees that, although the Representatives and the other underwriters may be required or choose to provide certain Regulation Best Interest and Form CRS disclosures to the undersigned in connection with the Offering, the Representatives and the other underwriters are not making a recommendation to the undersigned to enter into this agreement and nothing set forth in such disclosures is intended to suggest that the Representatives or any underwriter is making such a recommendation.
A-4
If (i)(a) prior to the execution of the Underwriting Agreement, the Company notifies the Representatives in writing that it does not intend to proceed with the Offering or (b) prior to the execution of the Underwriting Agreement, the Representatives notify the Company in writing that the underwriters do not intend to proceed with the Offering, (ii) the Underwriting Agreement is not executed by March 31, 2027 (provided, however, that the undersigned agrees that this agreement shall be automatically extended by three months if the Company provides written notice to the undersigned that the Company is still pursuing the Offering), (iii) the Underwriting Agreement (other than the provisions thereof which survive termination) shall terminate or be terminated for any reason prior to payment for and delivery of any Shares to be sold thereunder, or (iv) the registration statement filed with the Securities and Exchange Commission in connection with the Offering is withdrawn, then this agreement shall immediately be terminated and the undersigned shall automatically be released from all of his, her or its obligations under this agreement.
The undersigned hereby represents and warrants that the undersigned has full power, capacity and authority to enter into this agreement. This agreement is irrevocable and will be binding on the undersigned and the successors, heirs, personal representatives and assigns of the undersigned. This agreement may be delivered via facsimile, electronic mail (including pdf or any electronic signature complying with the U.S. federal ESIGN Act of 2000, e.g., www.docusign.com or www.echosign.com) or other transmission method and any counterpart so delivered shall be deemed to have been duly and validly delivered and be valid and effective for all purposes.
This agreement shall be governed by, and construed in accordance with, the laws of the State of New York.
A-5
| IF A NATURAL PERSON: | IF AN ENTITY OR TRUST: | |||||||
| By: |
|
| ||||||
|
|
(Please print complete name of entity) | |||||||
| (Duly authorized signature) | ||||||||
| Name: |
|
By: |
| |||||
|
|
| |||||||
| (Please print full name) | (Duly authorized signature) | |||||||
| Name: |
| |||||||
| (Please print full name) | ||||||||
| Title: |
| |||||||
| (Please print full title) | ||||||||
| Address: | Address: | |||||||
|
|
| |||||||
|
|
| |||||||
| E-mail: |
|
E-mail: |
| |||||
A-6
Annex A
Certain Defined Terms
Used in Lock-up Agreement
For purposes of the agreement to which this Annex A is attached and of which it is made a part:
| • | “Affiliate” shall have the meaning set forth in Rule 405 under the Securities Act. |
| • | “Call Equivalent Position” shall have the meaning set forth in Rule 16a-1(b) under the Exchange Act. |
| • | “Change of Control” means the consummation of any bona fide third party tender offer, merger, amalgamation, consolidation or other similar transaction following the completion of the Offering and approved by the Board of Directors of the Company, the result of which is that any “person” (as defined in Section 13(d)(3) of the Exchange Act), or group of persons, other than the Company or its subsidiaries, becomes the beneficial owner (as defined in Rules 13d-3 and 13d-5 of the Exchange Act) of more than 50% of the total voting power of the voting stock of the Company. |
| • | “Exchange Act” shall mean the Securities Exchange Act of 1934, as amended. |
| • | “Family Member” shall mean the spouse of the undersigned, an immediate family member of the undersigned or an immediate family member of the undersigned’s spouse, in each case living in the undersigned’s household or whose principal residence is the undersigned’s household (regardless of whether such spouse or family member may at the time be living elsewhere due to educational activities, health care treatment, military service, temporary internship or employment or otherwise). “Immediate family member” as used above shall have the meaning set forth in Rule 16a-1(e) under the Exchange Act. |
| • | “Lock-up Period” shall mean the period beginning on the date hereof and continuing through the close of trading on the date that is 180 days after the date of the Prospectus (as defined in the Underwriting Agreement). |
| • | “Put Equivalent Position” shall have the meaning set forth in Rule 16a-1(h) under the Exchange Act. |
| • | “Related Securities” shall mean any options or warrants or other rights to acquire Shares or any securities exchangeable or exercisable for or convertible into Shares, or to acquire other securities or rights ultimately exchangeable or exercisable for or convertible into Shares. |
| • | “Securities Act” shall mean the Securities Act of 1933, as amended. |
| • | “Sell or Offer to Sell” shall mean to: |
sell, offer to sell, contract to sell or lend,
effect any short sale or establish or increase a Put Equivalent Position or liquidate or decrease any Call Equivalent Position,
pledge, hypothecate or grant any security interest in, or
in any other way transfer or dispose of,
in each case whether effected directly or indirectly.
A-7
| • | “Swap” shall mean any swap, hedge or similar arrangement or agreement that transfers, in whole or in part, the economic risk of ownership of Shares or Related Securities, regardless of whether any such transaction is to be settled in securities, in cash or otherwise. |
Capitalized terms not defined in this Annex A shall have the meanings given to them in the body of this agreement.
A-8
Exhibit 3.1
AMENDED AND RESTATED CERTIFICATE OF INCORPORATION
OF
ELECTRA THERAPEUTICS, INC.
(Pursuant to Sections 242 and 245 of the
General Corporation Law of the State of Delaware)
ELECTRA THERAPEUTICS, INC., a corporation organized and existing under and by virtue of the provisions of the General Corporation Law of the State of Delaware (the “General Corporation Law”),
DOES HEREBY CERTIFY:
FIRST: That the name of this corporation is ELECTRA THERAPEUTICS, INC. and that this corporation was originally incorporated pursuant to the General Corporation Law on October 11, 2018.
SECOND: That the Board of Directors of this corporation duly adopted resolutions proposing to amend and restate the Certificate of Incorporation of this corporation, declaring said amendment and restatement to be advisable and in the best interests of this corporation and its stockholders, and authorizing the appropriate officers of this corporation to solicit the consent of the stockholders therefor, which resolution setting forth the proposed amendment and restatement is as follows:
RESOLVED, that the Certificate of Incorporation of this corporation be amended and restated in its entirety as follows:
ARTICLE I
The name of this corporation is Electra Therapeutics, Inc.
ARTICLE II
The address of the registered office of this corporation in the State of Delaware is 3500 South Dupont Highway, in the City of Dover, County of Kent, 19901. The name of its registered agent at such address is Incorporating Services, Ltd.
ARTICLE III
The nature of the business or purposes to be conducted or promoted is to engage in any lawful act or activity for which corporations may be organized under the General Corporation Law.
ARTICLE IV
A. Authorization of Stock. This corporation is authorized to issue two classes of stock to be designated, respectively, common stock and preferred stock. The total number of shares that this corporation is authorized to issue is 130,114,361. The total number of shares of common stock authorized to be issued is 71,000,000, par value $0.0001 per share (the “Common Stock”). The total number of shares of preferred stock authorized to be issued is 59,114,361 (the “Preferred Stock”), of which 22,601,626 shares, par value $0.0001 per share, are designated as “Series A Preferred Stock”, 9,309,492 shares, par value $0.0001 per share, are designated as “Series B Preferred Stock” and 27,203,243 shares, par value $0.001 per share, are designated as “Series C Preferred Stock.”
B. Rights, Preferences and Restrictions of Preferred Stock. The rights, preferences, privileges and restrictions granted to and imposed on the Preferred Stock are as set forth below in this Article IV(B).
1. Dividend Provisions.
(a) The holders of shares of Preferred Stock shall be entitled to receive dividends on a pari passu basis, out of any assets legally available therefor, prior and in preference to any declaration or payment of any dividend (payable other than in Common Stock or other securities and rights convertible into or entitling the holder thereof to receive, directly or indirectly, additional shares of Common Stock) on the Common Stock of this corporation at a rate of 8.0% of the Original Issue Price (as defined below) per annum, payable when, as and if declared by this corporation’s board of directors (the “Board of Directors”). Such dividends shall not be cumulative. The holders of the outstanding Preferred Stock can waive any dividend preference that such holders shall be entitled to receive under this Section 1 upon the affirmative vote or written consent of the holders of at least seventy percent (70%) of the shares of Series B Preferred Stock and Series C Preferred Stock then outstanding (voting together as a single class and not as separate series, and on an as-converted basis) (the “Requisite Holders”).
(b) After payment of such dividends, any additional dividends or distributions shall be distributed among all holders of Common Stock and Preferred Stock in proportion to the number of shares of Common Stock that would be held by each such holder if all shares of Preferred Stock were converted to Common Stock at the then effective Conversion Rate (as defined below). For this purpose, each holder of shares of Preferred Stock is to be treated as holding the greatest whole number of shares of Preferred Stock held by such holder pursuant to Section 4.
(c) Whenever a dividend provided for in this Section 1 will be payable in property other than cash, the value of such dividend will be deemed to be the fair market value of such property as determined in good faith by the Board of Directors, including the approval of a majority of the Preferred Directors who are then seated (the “Requisite Preferred Director Vote”).
(d) Notwithstanding the foregoing, in the event that this corporation declares a Special Dividend (as defined below), the proceeds of such Special Dividend shall be paid in accordance with Section 2 below and not this Section 1. A “Special Dividend” shall mean the payment of a dividend or other distribution of funds of this corporation to its stockholders where such funds were obtained from or are to be paid in connection with (i) the sale by this corporation of its equity securities (other than in connection with an initial public offering of this
2
corporation’s Common Stock) or incurrence of indebtedness for borrowed money outside of the ordinary course of business; (ii) the sale, lease, transfer, license or other disposition, in a single transaction or series of related transactions, by this corporation or any subsidiary of this corporation of a material portion of this corporation’s and its subsidiaries’ assets taken as a whole; or (iii) any other material transaction or series of related transactions that collectively would be material outside of the ordinary course of business; provided, however, that a Special Dividend shall not include (A) the distribution of funds or assets resulting from a Liquidation Event (as defined below) or (B) the distribution of funds or assets from this corporation’s cash flow from operation of this corporation’s business in the ordinary course.
2. Liquidation Preference.
(a) In the event of any Liquidation Event (as defined below), either voluntary or involuntary, or any Special Dividend, the holders of each series of Preferred Stock shall be entitled to receive, on a pari passu basis, out of the proceeds or assets of this corporation available for distribution to its stockholders (the “Proceeds”), prior and in preference to any distribution of the Proceeds of such Liquidation Event or Special Dividend to the holders of Common Stock by reason of their ownership thereof, an amount per share equal to the sum of the applicable Original Issue Price (as defined below) for such series of Preferred Stock, plus declared but unpaid dividends on such share (“Liquidation Amount”). If, upon the occurrence of such event, the Proceeds thus distributed among the holders of the Preferred Stock shall be insufficient to permit the payment to such holders of the full aforesaid preferential amounts, then the entire Proceeds legally available for distribution shall be distributed ratably among the holders of the Preferred Stock in proportion to the full preferential amount that each such holder is otherwise entitled to receive under this subsection (a). For purposes of this Amended and Restated Certificate of Incorporation (this “Restated Certificate of Incorporation”), “Original Issue Price” shall mean $1.5375 per share for each share of the Series A Preferred Stock, $9.0251 per share for each share of Series B Preferred Stock and $6.7688 per share for each share of Series C Preferred Stock (in each case, as adjusted for any stock splits, stock dividends, combinations, subdivisions, recapitalizations or the like with respect to such series of Preferred Stock).
(b) Upon completion of the distribution required by subsection (a) of this Section 2, all of the remaining Proceeds available for distribution to stockholders shall be distributed among the holders of Common Stock pro rata based on the number of shares of Common Stock held by each.
(c) Notwithstanding the above, for purposes of determining the amount each holder of shares of Preferred Stock is entitled to receive with respect to a Liquidation Event or Special Dividend, each such holder of shares of a series of Preferred Stock shall be deemed to have converted (regardless of whether such holder actually converted) such holder’s shares of such series into shares of Common Stock immediately prior to the Liquidation Event or Special Dividend if, as a result of an actual conversion, such holder would receive, in the aggregate, an amount greater than the amount that would be distributed to such holder if such holder did not convert such series of Preferred Stock into shares of Common Stock. If any such holder shall be deemed to have converted shares of Preferred Stock into Common Stock pursuant to this paragraph, then such holder shall not be entitled to receive any distribution that would otherwise be made to holders of Preferred Stock that have not converted (or have not been deemed to have
3
converted) into shares of Common Stock. In the event of more than one distribution pursuant to this Section 2, including distributions of Special Dividends and any proceeds of a Liquidation Event, such distributions shall be allocated among the holders of capital stock of this corporation in accordance with this Sections 2(a) and 2(b) after taking into account payment of the prior distributions so as to reach the result that would have occurred if all such proceeds had been distributed in a single event (for the purposes of example only, if this corporation first paid a Special Dividend of $10 million and subsequently paid a second Special Dividend of $20 million, the second Special Dividend would be distributed so as to equal the same results as if a single $30 million distribution had been made under this Section 2).
(d) (i) For purposes of this Section 2, a “Liquidation Event” shall include (A) the closing of the sale, lease, transfer, exclusive license or other disposition of all or substantially all of this corporation’s assets, (B) the consummation of the merger or consolidation of this corporation with or into another entity (except a merger or consolidation in which the holders of capital stock of this corporation immediately prior to such merger or consolidation continue to hold a majority of the voting power of the capital stock of this corporation or the surviving or acquiring entity immediately following such merger or consolidation in substantially the same proportions, and with substantially the same terms, as held immediately prior to such merger or consolidation, but expressly including such merger or consolidation with a publicly traded “special purpose acquisition company” or subsidiary of such company (collectively, a “SPAC”) to the extent such merger or consolidation with a SPAC is not a Qualifying SPAC Transaction (as defined below)), (C) the closing of the transfer (whether by merger, consolidation or otherwise), in one transaction or a series of related transactions, to a person or group of affiliated persons (other than an underwriter of this corporation’s securities), of this corporation’s securities if, after such closing, such person or group of affiliated persons would hold fifty percent (50%) or more of the then outstanding voting stock of this corporation (or the surviving or acquiring entity) or (D) a liquidation, dissolution or winding up of this corporation; provided, however, that a transaction shall not constitute a Liquidation Event if its sole purpose is to change the state of this corporation’s incorporation or to create a holding company that will be owned in substantially the same proportions by the persons who held this corporation’s securities immediately prior to such transaction. Notwithstanding the prior sentence, the sale by this corporation of its equity securities in a bona fide financing transaction conducted in accordance with Section 6 of this Article IV(B) shall not be deemed a “Liquidation Event.” The treatment of any particular transaction or series of related transactions as a Liquidation Event may be waived by the vote or written consent of the Requisite Holders.
(ii) In any Liquidation Event or Special Dividend, if Proceeds received by this corporation or its stockholders is other than cash, its value will be deemed its fair market value. Any securities shall be valued as follows:
(A) Securities not subject to investment letter or other similar restrictions on free marketability covered by (B) below:
(1) If traded on a securities exchange, the value shall be deemed to be the average of the closing prices of the securities on such exchange over the twenty (20) trading-day period ending three (3) trading days prior to the closing of the Liquidation Event or Special Dividend;
4
(2) If actively traded over-the-counter, the value shall be deemed to be the average of the closing bid or sale prices (whichever is applicable) over the twenty (20) trading-day period ending three (3) trading days prior to the closing of the Liquidation Event or Special Dividend; and
(3) If there is no active public market, the value shall be the fair market value thereof, as determined in good faith by Requisite Preferred Director Vote.
(B) The method of valuation of securities subject to investment letter or other restrictions on free marketability (other than restrictions arising solely by virtue of a stockholder’s status as an affiliate or former affiliate) shall be to make an appropriate discount from the market value determined as above in (A)(1), (A)(2) or (A)(3) to reflect the approximate fair market value thereof, as determined in good faith by Requisite Preferred Director Vote.
(C) The foregoing methods for valuing non-cash consideration to be distributed in connection with a Liquidation Event or Special Dividend shall, with the appropriate approval of the definitive agreements governing such Liquidation Event or Special Dividend by the stockholders under the General Corporation Law and Section 6 of this Article IV(B), be superseded by the determination of such value set forth in the definitive agreements governing such Liquidation Event or Special Dividend, as applicable.
(iii) In the event the requirements of this Section 2 are not complied with, this corporation shall forthwith either:
(A) cause the closing of such Liquidation Event or Special Dividend to be postponed until such time as the requirements of this Section 2 have been complied with; or
(B) cancel such transaction, in which event the rights, preferences and privileges of the holders of the Preferred Stock shall revert to and be the same as such rights, preferences and privileges existing immediately prior to the date of the first notice referred to in subsection 2(d)(iv) hereof.
(iv) This corporation shall give each holder of record of Preferred Stock written notice of such impending Liquidation Event or Special Dividend not later than twenty (20) days prior to the stockholders’ meeting called to approve such transaction, or twenty (20) days prior to the closing of such transaction, whichever is earlier, and shall also notify such holders in writing of the final approval of such transaction. The first of such notices shall describe the material terms and conditions of the impending transaction and the provisions of this Section 2, and this corporation shall thereafter give such holders prompt notice of any material changes. The transaction shall in no event take place sooner than twenty (20) days after this corporation has given the first notice provided for herein or sooner than ten (10) days after this corporation has given notice of any material changes provided for herein; provided, however, that subject to compliance with the General Corporation Law such periods may be shortened upon the written consent of the Requisite Holders.
5
(e) In the event of a Liquidation Event referred to in Section 2(d)(i)(A), if this corporation does not effect a dissolution under the General Corporation Law within ninety (90) days after such Liquidation Event, then (i) this corporation shall send a written notice to each holder of Preferred Stock no later than the ninetieth (90th) day after the Liquidation Event advising such holders of their right (and the requirements to be met to secure such right) pursuant to the terms of the following clause (ii) to require the redemption of such shares of Preferred Stock, and (iii) if the Requisite Holders so request in a written instrument delivered to this corporation not later than one hundred twenty (120) days after such Liquidation Event, this corporation shall use the consideration received for such Liquidation Event (net of any retained liabilities associated with the assets sold or technology licensed, as determined in good faith by Requisite Preferred Director Vote), together with any other assets for distribution to stockholders, all to the extent permitted by Delaware law governing distributions to stockholders (the “Available Proceeds”), on the one hundred fiftieth (150th) day after such Liquidation Event (the “LE Redemption Date”), to redeem all outstanding shares of Preferred Stock at a price per share equal to the Liquidation Amount; provided, that if the definitive agreements governing such Liquidation Event contain contingent indemnification obligations on the part of this corporation and prohibit this corporation from distributing all or a portion of the Available Proceeds while such indemnification obligations remain outstanding, then the LE Redemption Date shall automatically be extended to the date that is ten (10) business days following the date on which such prohibition expires. Notwithstanding the foregoing, in the event of a redemption pursuant to the preceding sentence, if the Available Proceeds are not sufficient to redeem all outstanding shares of Preferred Stock, this corporation shall redeem a pro rata portion of each holder’s shares of Preferred Stock to the fullest extent of such Available Proceeds, based on the respective amounts which would otherwise be payable in respect of the shares to be redeemed if the Available Proceeds were sufficient to redeem all such shares, and shall redeem the remaining shares as soon as it may lawfully do so under Delaware law governing distributions to stockholders. Prior to the distribution or redemption provided for in this Section 2(e), this corporation shall not expend or dissipate the consideration received for such Liquidation Event, except to discharge expenses incurred in connection with such Liquidation Event or in the ordinary course of business.
(f) Following an election by the holders of Preferred Stock to demand redemption as provided in Section 2(e), this corporation shall promptly send written notice of the redemption pursuant to Section 2(e) (the “Redemption Notice”) to each holder of record of Preferred Stock stating (i) the number of shares of each series of Preferred Stock held by such holder as of the date of such election; (ii) the redemption date and the price at which shares of each series of Preferred Stock will be redeemed (the “Redemption Price”); (iii) the date on which the holder’s right to convert such shares terminates (as determined in accordance with Section 4(a)); and (iv) for holders of shares in certificated form, that the holder is to surrender to this corporation, in the manner and at the place designated, his, her or its certificate or certificates representing the shares of Preferred Stock to be redeemed.
(g) On or before the redemption date, each holder of shares of Preferred Stock, unless such holder has exercised his, her or its right to convert such shares as provided in Section 4, shall, if a holder of shares in certificated form, surrender the certificate or certificates representing such shares (or, if such registered holder alleges that such certificate has been lost, stolen or destroyed, a lost certificate affidavit and agreement reasonably acceptable to this corporation to indemnify this corporation against any claim that may be made against this corporation on account of the alleged loss, theft or destruction of such certificate) to this corporation, in the manner and at the place designated in the Redemption Notice, and thereupon the Redemption Price for such shares shall be payable to the order of the person whose name appears on such certificate or certificates as the owner thereof.
6
(h) If the Redemption Notice shall have been duly given, and if on the redemption date the Redemption Price payable upon redemption of the shares of Preferred Stock is paid or tendered for payment or deposited with an independent payment agent so as to be available therefor in a timely manner, then notwithstanding that any certificates evidencing any of the shares of Preferred Stock so called for redemption shall not have been surrendered, dividends with respect to such shares of Preferred Stock shall cease to accrue after the redemption date and all rights with respect to such shares shall forthwith after the redemption date terminate, except only the right of the holders to receive the Redemption Price without interest upon surrender of any such certificate or certificates therefor.
(i) Allocation of Contingent Consideration. In the event of a Liquidation Event pursuant to subsection 2(d)(i), if any portion of the consideration payable to the stockholders of this corporation is placed into escrow and/or is payable to the stockholders of this corporation subject to contingencies (“Additional Consideration”), the definitive agreement with respect to such deemed Liquidation Event shall provide that (i) the portion of such consideration that is not Additional Consideration (such portion, the “Initial Consideration”) shall be allocated among the holders of capital stock of this corporation in accordance with subsections 2(a) and 2(b) as if the Initial Consideration were the only consideration payable in connection with such deemed Liquidation Event and (ii) any Additional Consideration that becomes payable to the stockholders of this corporation upon release from escrow or satisfaction of contingencies shall be allocated among the holders of capital stock of this corporation in accordance with subsections 2(a) and 2(b) after taking into account the previous payment of the Initial Consideration as part of the same transaction. For the purposes of this subsection 2(i), consideration placed into escrow or retained as a holdback to be available for satisfaction of indemnification or similar obligations in connection with such deemed Liquidation Event shall be deemed to be Additional Consideration.
3. Redemption. The Preferred Stock is not redeemable at the option of the holder thereof except for redemption pursuant to Section 2(e).
4. Conversion. The holders of the Preferred Stock shall have conversion rights as follows (the “Conversion Rights”):
(a) Right to Convert. Subject to Section 4A.1., each share of Preferred Stock shall be convertible, at the option of the holder thereof, at any time after the date of issuance of such share, at the office of this corporation or any transfer agent for such stock, into such number of fully paid and nonassessable shares of Common Stock as is determined by dividing the applicable Original Issue Price for such series by the applicable Conversion Price (as defined below) for such series (the conversion rate for a series of Preferred Stock into Common Stock is referred to herein as the “Conversion Rate” for such series), determined as hereafter provided, in effect on the date the certificate is surrendered for conversion. The initial “Conversion Price” per share for each series of Preferred Stock shall be the Original Issue Price applicable to such series; provided, however, that the Conversion Price for the Preferred Stock shall be subject to adjustment as set forth in subsection 4(d).
7
(b) Automatic Conversion. Each share of Preferred Stock shall automatically be converted into shares of Common Stock at the Conversion Rate at the time in effect for such series of Preferred Stock immediately upon the earlier of (i) immediately prior to the closing of this corporation’s sale of its Common Stock to the public at a price of at least $6.7688 per share (subject to appropriate adjustment in the event of any stock dividend, stock split, combination or other similar recapitalization with respect to the Common Stock), in a firm commitment underwritten public offering pursuant to an effective registration statement on Form S-1 under the Securities Act of 1933, as amended, that results in at least $100,000,000 of gross proceeds and in connection with such offering the Common Stock is listed for trading on the Nasdaq Stock Market, the New York Stock Exchange or another exchange or marketplace approved by the Board of Directors, (ii) this corporation’s completion of a transaction or series of related transactions (by merger, or consolidation, share exchange or otherwise) with a SPAC in which the common stock or share capital of the SPAC or its successor or ultimate parent entity (collectively, the “Listed Entity”) is listed for trading on the Nasdaq Stock Market’s National Market, the New York Stock Exchange or another exchange or marketplace approved by the Board of Directors and, in connection with which (a) this corporation is valued in such SPAC transaction such that each share of Series C Preferred Stock will be exchanged for a share of capital stock of the Listed Entity with a per share value that is at least $6.7688 (subject to appropriate adjustment in the event of any stock dividend, stock split, combination or other similar recapitalization with respect to the Common Stock) and (b) the Listed Entity, upon closing of such transaction, has unrestricted cash of an aggregate of at least $100,000,000 (the foregoing, a “Qualified SPAC Transaction”), (iii) the effectiveness of the registration statement in connection with the initial listing of the Common Stock (or other equity securities of this corporation) on the Nasdaq Stock Market’s National Market, the New York Stock Exchange or another exchange or marketplace approved by the Board of Directors by means of an effective registration statement filed by this corporation with the Securities and Exchange Commission, without a related underwritten offering of such Common Stock (or other equity securities) or (iv) the date, or the occurrence of an event, specified by vote or written consent or agreement of the Requisite Holders.
(c) Mechanics of Conversion. Before any holder of Preferred Stock shall be entitled to voluntarily convert the same into shares of Common Stock, such holder shall surrender the certificate or certificates therefor, duly endorsed, at the office of this corporation or of any transfer agent for the Preferred Stock, and shall give written notice to this corporation at its principal corporate office, of the election to convert the same and shall state therein the name or names in which the certificate or certificates for shares of Common Stock are to be issued. This corporation shall, as soon as practicable thereafter, issue and deliver at such office to such holder of Preferred Stock, or to the nominee or nominees of such holder, a certificate or certificates for the number of shares of Common Stock to which such holder shall be entitled as aforesaid. Such conversion shall be deemed to have been made immediately prior to the close of business on the date set forth for conversion in the written notice of the election to convert irrespective of the surrender of the shares of Preferred Stock to be converted, and the person or persons entitled to receive the shares of Common Stock issuable upon such conversion shall be treated for all purposes as the record holder or holders of such shares of Common Stock as of
8
such date. If the conversion is in connection with an underwritten offering of securities registered pursuant to the Securities Act of 1933, as amended, the conversion may, at the option of any holder tendering Preferred Stock for conversion, be conditioned upon the closing with the underwriters of the sale of securities pursuant to such offering, in which event the persons entitled to receive the Common Stock upon conversion of the Preferred Stock shall not be deemed to have converted such Preferred Stock until immediately prior to the closing of such sale of securities. If the conversion is in connection with Automatic Conversion provisions of subsection 4(b)(ii) above, such conversion shall be deemed to have been made immediately prior to the close of business on the conversion date described in the stockholder consent approving such conversion, and the persons entitled to receive shares of Common Stock issuable upon such conversion shall be treated for all purposes as the record holders of such shares of Common Stock as of such date.
(d) Conversion Price Adjustments of Preferred Stock for Certain Dilutive Issuances, Splits and Combinations. The Conversion Price of the Preferred Stock shall be subject to adjustment from time to time as follows:
(i) (A) If this corporation shall issue, on or after the date on which the first share of Series C Preferred is issued (the “Series C Original Issue Date”), any Additional Stock (as defined below) without consideration or for a consideration per share less than the Conversion Price applicable to a series of Preferred Stock in effect immediately prior to the issuance of such Additional Stock, the Conversion Price for such series in effect immediately prior to each such issuance shall forthwith (except as otherwise provided in this clause (i)) be adjusted to a price (calculated to the nearest one-thousandth of a cent) determined by multiplying such Conversion Price by a fraction, the numerator of which shall be the number of shares of Common Stock Outstanding (as defined below) immediately prior to such issuance plus the number of shares of Common Stock that the aggregate consideration received by this corporation for such issuance would purchase at such Conversion Price; and the denominator of which shall be the number of shares of Common Stock Outstanding (as defined below) immediately prior to such issuance plus the number of shares of such Additional Stock. For purposes of this Section 4(d)(i)(A), the term “Common Stock Outstanding” shall mean and include the following: (1) outstanding Common Stock, (2) Common Stock issuable upon conversion of outstanding Preferred Stock, (3) Common Stock issuable upon exercise of outstanding stock options and (4) Common Stock issuable upon exercise (and, in the case of warrants to purchase Preferred Stock, conversion) of outstanding warrants. Shares described in (1) through (4) above shall be included whether vested or unvested, whether contingent or non-contingent and whether exercisable or not yet exercisable. In the event that this corporation issues or sells, or is deemed to have issued or sold, shares of Additional Stock that results in an adjustment to a Conversion Price pursuant to the provisions of this Section 4(d) (the “First Dilutive Issuance”), and this corporation then issues or sells, or is deemed to have issued or sold, shares of Additional Stock in one or more subsequent issuances other than the First Dilutive Issuance that would result in further adjustment to a Conversion Price (each, a “Subsequent Dilutive Issuance”) pursuant to the same instruments as the First Dilutive Issuance, then and in each such case upon a Subsequent Dilutive Issuance the applicable Conversion Price for each series of Preferred Stock shall be reduced to the applicable Conversion Price that would have been in effect had the First Dilutive Issuance and each Subsequent Dilutive Issuance all occurred on the closing date of the First Dilutive Issuance.
9
(B) No adjustment of the Conversion Price for the Preferred Stock shall be made in an amount less than one-tenth of one cent per share. Except to the limited extent provided for in subsections (E)(3) and (E)(4), no adjustment of such Conversion Price pursuant to this subsection 4(d)(i) shall have the effect of increasing the Conversion Price above the Conversion Price in effect immediately prior to such adjustment.
(C) In the case of the issuance of Additional Stock for cash, the consideration shall be deemed to be the amount of cash paid therefor before deducting any reasonable discounts, commissions or other expenses allowed, paid or incurred by this corporation for any underwriting or otherwise in connection with the issuance and sale thereof.
(D) In the case of the issuance of the Additional Stock for a consideration in whole or in part other than cash, the consideration other than cash shall be deemed to be the fair market value thereof as determined in good faith by the Board of Directors irrespective of any accounting treatment.
(E) In the case of the issuance of options to purchase or rights to subscribe for Common Stock, securities by their terms convertible into or exchangeable for Common Stock or options to purchase or rights to subscribe for such convertible or exchangeable securities, the following provisions shall apply for purposes of determining the number of shares of Additional Stock issued and the consideration paid therefor:
(1) The aggregate maximum number of shares of Common Stock deliverable upon exercise (assuming the satisfaction of any conditions to exercisability, including without limitation, the passage of time, but without taking into account potential antidilution adjustments) of such options to purchase or rights to subscribe for Common Stock shall be deemed to have been issued at the time such options or rights were issued and for a consideration equal to the consideration (determined in the manner provided in subsections 4(d)(i)(C) and (d)(i)(D)), if any, received by this corporation upon the issuance of such options or rights plus the minimum exercise price provided in such options or rights (without taking into account potential antidilution adjustments) for the Common Stock covered thereby.
(2) The aggregate maximum number of shares of Common Stock deliverable upon conversion of, or in exchange (assuming the satisfaction of any conditions to convertibility or exchangeability, including, without limitation, the passage of time, but without taking into account potential antidilution adjustments) for, any such convertible or exchangeable securities or upon the exercise of options to purchase or rights to subscribe for such convertible or exchangeable securities and subsequent conversion or exchange thereof shall be deemed to have been issued at the time such securities were issued or such options or rights were issued and for a consideration equal to the consideration, if any, received by this corporation for any such securities and related options or rights (excluding any cash received on account of accrued interest or accrued dividends), plus the minimum additional consideration, if any, to be received by this corporation (without taking into account potential antidilution adjustments) upon the conversion or exchange of such securities or the exercise of any related options or rights (the consideration in each case to be determined in the manner provided in subsections 4(d)(i)(C) and (d)(i)(D)). If the number of shares of Common Stock issuable upon the exercise, conversion and/or exchange of any convertible or exchangeable securities, or the consideration payable to the
10
corporation upon such exercise, conversion and/or exchange, cannot be calculated at all at the time such convertible or exchangeable securities is issued or amended, any adjustment to the Conversion Price of a series of Preferred Stock that would result under the terms of this Section 4(d)(i)(E) at the time of such issuance or amendment shall instead be effected at the time such number of shares and/or amount of consideration is first calculable (even if subject to subsequent adjustments), assuming for purposes of calculating such adjustment to the Conversion Price for such series of Preferred Stock that such issuance or amendment took place at the time such calculation can first be made. In the event any convertible or exchangeable securities contains alternative conversion terms, such as a cap on the valuation of the corporation at which such conversion will be effected, or circumstances where the convertible or exchangeable securities may be repaid in lieu of conversion, then the number of shares of Common Stock issuable upon the exercise, conversion and/or exchange of such convertible or exchangeable securities shall be deemed not calculable until such time as the applicable conversion terms are determined.
(3) In the event of any change in the number of shares of Common Stock deliverable or in the consideration payable to this corporation upon exercise of such options or rights or upon conversion of or in exchange for such convertible or exchangeable securities, the Conversion Price of the Preferred Stock, to the extent in any way affected by or computed using such options, rights or securities, shall be recomputed to reflect such change, but no further adjustment shall be made for the actual issuance of Common Stock or any payment of such consideration upon the exercise of any such options or rights or the conversion or exchange of such securities.
(4) Upon the expiration of any such options or rights, the termination of any such rights to convert or exchange or the expiration of any options or rights related to such convertible or exchangeable securities, the Conversion Price of the Preferred Stock, to the extent in any way affected by or computed using such options, rights or securities or options or rights related to such securities, shall be recomputed to reflect the issuance of only the number of shares of Common Stock (and convertible or exchangeable securities that remain in effect) actually issued upon the exercise of such options or rights, upon the conversion or exchange of such securities or upon the exercise of the options or rights related to such securities.
(5) The number of shares of Additional Stock deemed issued and the consideration deemed paid therefor pursuant to subsections 4(d)(i)(E)(1) and (2) shall be appropriately adjusted to reflect any change, termination or expiration of the type described in either subsection 4(d)(i)(E)(3) or (4).
(ii) “Additional Stock” shall mean any shares of Common Stock issued (or deemed to have been issued pursuant to subsection 4(d)(i)(E)) by this corporation on or after the Series C Original Issue Date other than (“Excluded Securities”):
(A) Common Stock issued pursuant to a transaction described in subsection 4(d)(iii) hereof;
11
(B) Common Stock issued to employees, directors, consultants and other service providers for the primary purpose of soliciting or retaining their services pursuant to plans or agreements approved by the Board of Directors;
(C) Common Stock issued pursuant to an underwritten public offering;
(D) Common Stock issued pursuant to the conversion or exercise of convertible or exercisable securities outstanding on the Series C Original Issue Date, in each case provided such issuance is pursuant to the terms of such convertible or exercisable security;
(E) Common Stock issued as acquisition consideration in connection with a bona fide business acquisition by this corporation, whether by merger, consolidation, sale of assets, sale or exchange of stock or otherwise provided that such issuances are approved by Requisite Preferred Director Vote;
(F) Common Stock issued or deemed issued pursuant to subsection 4(d)(i)(E) as a result of a decrease in the Conversion Price of any series of Preferred Stock resulting from the operation of Section 4(d);
(G) Common Stock issued upon conversion of Preferred Stock;
(H) Shares issued pursuant to the Series C Preferred Stock Purchase Agreement, by and among this corporation and certain investors dated as of the date the first shares of Series C Preferred Stock are issued by this corporation (as amended from time to time, the “Purchase Agreement”);
(I) Common Stock issued pursuant to any equipment leasing arrangement or debt financing arrangement, which arrangement is approved by Requisite Preferred Director Vote; or
(J) Common Stock issued to suppliers or third party service providers in connection with the provision of goods or services or issued in connection with sponsored research, collaboration, technology license, development, OEM, marketing or other similar agreements or strategic partnerships, provided such transactions and such issuances are approved by Requisite Preferred Director Vote.
(iii) In the event this corporation should at any time or from time to time after the Series C Original Issue Date fix a record date for the effectuation of a split or subdivision of the outstanding shares of Common Stock or the determination of holders of Common Stock entitled to receive a dividend or other distribution payable in additional shares of Common Stock or other securities or rights convertible into, or entitling the holder thereof to receive directly or indirectly, additional shares of Common Stock (hereinafter referred to as “Common Stock Equivalents”) without payment of any consideration by such holder for the additional shares of Common Stock or the Common Stock Equivalents (including the additional shares of Common Stock issuable upon conversion or exercise thereof), then, as of such record
12
date (or the date of such dividend distribution, split or subdivision if no record date is fixed), the Conversion Price of the Preferred Stock shall be appropriately decreased so that the number of shares of Common Stock issuable on conversion of each share of such series shall be increased in proportion to such increase of the aggregate of shares of Common Stock outstanding and those issuable with respect to such Common Stock Equivalents with the number of shares issuable with respect to Common Stock Equivalents determined from time to time in the manner provided for deemed issuances in subsection 4(d)(i)(E).
(iv) If the number of shares of Common Stock outstanding at any time after the Series C Original Issue Date is decreased by a combination of the outstanding shares of Common Stock, then, following the record date of such combination, the Conversion Price for the Preferred Stock shall be appropriately increased so that the number of shares of Common Stock issuable on conversion of each share of such series shall be decreased in proportion to such decrease in outstanding shares.
(e) Other Distributions. In the event this corporation shall declare a distribution payable in securities of other persons, evidences of indebtedness issued by this corporation or other persons, assets (excluding cash dividends) or options or rights not referred to in subsection 4(d)(iii), then, in each such case for the purpose of this subsection 4(e), the holders of the Preferred Stock shall be entitled to a proportionate share of any such distribution as though they were the holders of the number of shares of Common Stock of this corporation into which their shares of Preferred Stock are convertible as of the record date fixed for the determination of the holders of Common Stock of this corporation entitled to receive such distribution.
(f) Recapitalizations. If at any time or from time to time there shall be a recapitalization of the Common Stock (other than a subdivision, combination or merger or sale of assets transaction provided for elsewhere in this Section 4 or in Section 2) provision shall be made so that the holders of the Preferred Stock shall thereafter be entitled to receive upon conversion of the Preferred Stock the number of shares of stock or other securities or property of this corporation or otherwise, to which a holder of Common Stock deliverable upon conversion would have been entitled on such recapitalization. In any such case, appropriate adjustment shall be made in the application of the provisions of this Section 4 with respect to the rights of the holders of the Preferred Stock after the recapitalization to the end that the provisions of this Section 4 (including adjustment of the Conversion Price then in effect and the number of shares purchasable upon conversion of the Preferred Stock) shall be applicable after that event as nearly equivalently as may be practicable.
(g) No Fractional Shares and Certificate as to Adjustments.
(i) No fractional shares shall be issued upon the conversion of any share or shares of the Preferred Stock and the aggregate number of shares of Common Stock to be issued to particular stockholders, shall be rounded down to the nearest whole share and this corporation shall pay in cash the fair market value of any fractional shares as of the time when entitlement to receive such fractional shares is determined. Whether or not fractional shares would be issuable upon such conversion shall be determined on the basis of the total number of shares of Preferred Stock the holder is at the time converting into Common Stock and the number of shares of Common Stock issuable upon such conversion.
13
(ii) Upon the occurrence of each adjustment or readjustment of the Conversion Price of Preferred Stock pursuant to this Section 4, this corporation, at its expense, shall promptly compute such adjustment or readjustment in accordance with the terms hereof and prepare and furnish to each holder of Preferred Stock a certificate setting forth such adjustment or readjustment and showing in detail the facts upon which such adjustment or readjustment is based. This corporation shall, upon the written request at any time of any holder of Preferred Stock, furnish or cause to be furnished to such holder a like certificate setting forth (A) such adjustment and readjustment, (B) the Conversion Price for such series of Preferred Stock at the time in effect, and (C) the number of shares of Common Stock and the amount, if any, of other property that at the time would be received upon the conversion of a share of Preferred Stock.
(h) Notices of Record Date. In the event of any taking by this corporation of a record of the holders of any class of securities for the purpose of determining the holders thereof who are entitled to receive any dividend (other than a cash dividend) or other distribution, this corporation shall mail to each holder of Preferred Stock, at least ten (10) days prior to the date specified therein, a notice specifying the date on which any such record is to be taken for the purpose of such dividend or distribution, and the amount and character of such dividend or distribution; provided, however, that subject to compliance with the General Corporation Law such notice period may be shortened upon the written consent of the Requisite Holders.
(i) Reservation of Stock Issuable Upon Conversion. This corporation shall at all times reserve and keep available out of its authorized but unissued shares of Common Stock, solely for the purpose of effecting the conversion of the shares of the Preferred Stock, such number of its shares of Common Stock as shall from time to time be sufficient to effect the conversion of all outstanding shares of the Preferred Stock; and if at any time the number of authorized but unissued shares of Common Stock shall not be sufficient to effect the conversion of all then outstanding shares of the Preferred Stock, in addition to such other remedies as shall be available to the holder of such Preferred Stock, this corporation will take such corporate action as may, in the opinion of its counsel, be necessary to increase its authorized but unissued shares of Common Stock to such number of shares as shall be sufficient for such purposes, including, without limitation, engaging in best efforts to obtain the requisite stockholder approval of any necessary amendment to this Restated Certificate of Incorporation.
(j) Waiver of Adjustment to Conversion Price. Notwithstanding anything herein to the contrary, any downward adjustment of the Conversion Price of any series of Preferred Stock may be waived, either prospectively or retroactively and either generally or in a particular instance, by the consent or vote of the holders of (i) solely with respect to the Conversion Price of the Series A Preferred Stock, a majority of the then outstanding shares of Series A Preferred Stock (voting together as a single class and on an as-converted basis); (ii) solely with respect to the Conversion Price of the Series B Preferred Stock a majority of the then outstanding shares of Series B Preferred Stock (voting together as a single class and on an as-converted basis); and (iii) solely with respect to the Conversion Price of the Series C Preferred Stock at least 60% of the then outstanding shares of Series C Preferred Stock (voting together as a single class and on an as-converted basis). Any such waiver shall bind all future holders of shares of such series of Preferred Stock.
14
4A. Special Mandatory Conversion.
4A.1. Trigger Event. In the event that any holder of shares of Series C Preferred Stock is a Defaulting Investor (as defined in the Purchase Agreement), then each Subject Share (as defined in the Purchase Agreement) held by such Defaulting Investor and its Affiliates shall automatically, and without further action on the part of such Defaulting Investor and its Affiliates, be converted into such number of fully paid and non-assessable shares of Common Stock as is determined by dividing the applicable Original Issue Price of such Subject Share by a price equal to ten times (10X) the applicable Conversion Price of such Subject Share in effect at the time of conversion, effective upon, subject to, and concurrently with, the consummation of the Second Tranche Closing. Such conversion is referred to as a “Special Mandatory Conversion.” For purposes of this Section 4A, “Affiliate” shall mean, with respect to any Investor (as defined in the Purchase Agreement), any person, entity or firm which, from time to time, directly or indirectly, controls, is controlled by, or is under common control with such Investor, including, without limitation, any general partner, managing member, officer, director or trustee of such Investor, or any venture capital fund or registered investment company now or hereafter existing that is controlled by one or more general partners, managing members or investment advisers of, or shares the same management company or investment adviser with, such Investor. Notwithstanding anything to the contrary, the optional conversion rights of the Series B Preferred Stock and Series C Preferred Stock set forth in Section 4(a) shall not be effective for any of the Investors or their Affiliates until the earliest of (i) the day immediately following the Second Tranche Closing Date, (ii) a Liquidation Event, (iii) the date the obligation to purchase the Second Tranche Shares (as defined in the Purchase Agreement) terminates in accordance with the terms of the Purchase Agreement, or (iv) such other time as specified by this corporation and the Requisite Investors (as defined in the Purchase Agreement) (such earlier date, the “Expiration Date”).
4A.2. Procedural Requirements. Upon a Special Mandatory Conversion, each Defaulting Investor and its Affiliates that hold Subject Shares to be converted pursuant to Section 4A.1 shall be sent written notice of such Special Mandatory Conversion and the place designated for mandatory conversion of all such Subject Shares pursuant to this Section 4A.2. Upon receipt of such notice, each Defaulting Investor and its Affiliates that hold such Subject Shares in certificated form shall surrender his, her or its certificate or certificates for all such shares (or, if such holder alleges that any such certificate has been lost, stolen or destroyed, a lost certificate affidavit and agreement reasonably acceptable to this corporation to indemnify this corporation against any claim that may be made against this corporation on account of the alleged loss, theft or destruction of such certificate) to this corporation at the place designated in such notice. If so required by this corporation, any certificates surrendered for conversion shall be endorsed or accompanied by written instrument or instruments of transfer, in form satisfactory to this corporation, duly executed by the registered holder or by his, her or its attorney duly authorized in writing. All rights with respect to the Subject Shares converted pursuant to Section 4A.1, including the rights, if any, to receive notices and vote (other than as a holder of Common Stock), will terminate at the time of the Special Mandatory Conversion (notwithstanding the failure of the holder or holders thereof to surrender any certificates for such shares at or prior to such time), except only the rights of the holders thereof, upon surrender of any certificate or certificates of such holders therefor (or lost certificate affidavit and agreement), to receive the items provided for in the next sentence of this Section 4A.2. As soon as practicable after the Special Mandatory Conversion and, if applicable, the surrender of any certificate or certificates (or lost certificate affidavit and agreement) for Subject Shares so converted, this corporation shall (a) issue and deliver to such holder, or to his, her or its nominees, a certificate or certificates for the number of full shares of Common Stock issuable on such conversion in accordance with the provisions hereof and (b) pay cash as provided in Section 4(g) in lieu of any fraction of a share of Common Stock otherwise issuable upon such conversion and the payment of any declared but unpaid dividends on the Subject Shares converted.
15
Such converted Subject Shares shall be retired and cancelled and may not be reissued as shares of such series, and this corporation may thereafter take such appropriate action (without the need for stockholder action) as may be necessary to reduce the authorized number of shares of Series B Preferred Stock and Series C Preferred Stock accordingly.
5. Voting Rights.
(a) General Voting Rights. The holder of each share of Preferred Stock shall have the right to one vote for each share of Common Stock into which such Preferred Stock could then be converted, and with respect to such vote, such holder shall have full voting rights and powers equal to the voting rights and powers of the holders of Common Stock, and shall be entitled, notwithstanding any provision hereof, to notice of any stockholders’ meeting in accordance with the Bylaws of this corporation, and except as provided by law or in subsection 5(b) below with respect to the election of directors by the separate class vote of the holders of Common Stock, shall be entitled to vote, together with holders of Common Stock, with respect to any question upon which holders of Common Stock have the right to vote. Fractional votes shall not, however, be permitted and any fractional voting rights available on an as-converted basis (after aggregating all shares into which shares of Preferred Stock held by each holder could be converted) shall be rounded to the nearest whole number (with one-half (1/2) being rounded upward).
(b) Voting for the Election of Directors. The holders of such shares of Series B Preferred Stock, exclusively and as a separate class, shall be entitled to elect two (2) directors of this corporation (the “Series B Directors”) at any election of directors and the holders of such shares of Series C Preferred Stock, exclusively and as a separate class, shall be entitled to elect one director of this corporation (the “Series C Director”, together with the Series B Directors, the “Preferred Directors”) at any election of directors. Any remaining directors of this corporation shall be elected by the holders of a majority of the then outstanding stock of the corporation (voting together as a single class and not as separate series, and on an as-converted basis) (the “Joint Directors”).
Notwithstanding the provisions of Section 223(a)(1) and 223(a)(2) of the General Corporation Law, any vacancy, including newly created directorships resulting from any increase in the authorized number of directors or amendment of this Restated Certificate of Incorporation, and vacancies created by removal or resignation of a director, may be filled by a majority of the directors then in office, though less than a quorum, or by a sole remaining director, and the directors so chosen shall hold office until the next annual election and until their successors are duly elected and qualified, unless sooner displaced; provided, however, that where such vacancy occurs among the directors elected by the holders of a class or series of stock, the holders of shares of such class or series may override the Board of Directors’ action to fill such vacancy by (i) voting for their own designee to fill such vacancy at a meeting of this corporation’s stockholders or (ii) written consent, if the consenting stockholders hold a sufficient number of shares to elect their designee at a meeting of the stockholders.
16
Any director may be removed during his or her term of office, either with or without cause, by, and only by, the affirmative vote of the holders of the shares of the class or series of stock entitled to elect such director or directors, given either at a special meeting of such stockholders duly called for that purpose or pursuant to a written consent of stockholders, and any vacancy thereby created may be filled by the holders of that class or series of stock represented at the meeting or pursuant to written consent.
6. Protective Provisions.
(a) Preferred Stock Protective Provisions. So long as 25% of the shares of Preferred Stock originally issued (as adjusted for any stock splits, stock dividends, combinations, subdivisions, recapitalizations or the like with respect to such Preferred Stock) remains outstanding, this corporation shall not, either directly or indirectly by amendment, merger, domestication, transfer, continuance, reorganization, recapitalization, reclassification, waiver, statutory conversion, consolidation or otherwise) effect any of the following acts or transactions without (in addition to any other vote required by law or this Restated Certificate of Incorporation) first obtaining the approval by vote or written consent, as provided by law, of the Requisite Holders (and any such act or transaction that has not been approved by such consent or vote prior to such act or transaction being effected shall be null and void ab initio, and of no force or effect):
(i) consummate a Liquidation Event;
(ii) amend, alter or repeal any provision of this Restated Certificate of Incorporation or Bylaws;
(iii) increase or decrease (other than by redemption or conversion) the total number of authorized shares of Common Stock or Preferred Stock or designated shares of any series of Preferred Stock;
(iv) (A) create or adopt, any equity (or equity-linked) compensation plan; or (B) materially amend any such plan, which shall include increasing the number of shares authorized for issuance thereunder;
(v) unless approved by Requisite Preferred Director Vote, incur any indebtedness for borrowed money, including but not limited to obligations and contingent obligations under guarantees, or permit any subsidiary to take any such action with respect to any debt security lien, security interest or other indebtedness for borrowed money, if the aggregate indebtedness of this corporation and its subsidiaries would exceed one million dollars ($1,000,000.00) (other than equipment leases, bank lines of credit, letters of credit or trade payables, in each case, incurred in the ordinary course), or create any lien or security interest (except for purchase money liens or statutory liens of landlords, mechanics, materialmen, workmen, warehousemen and other similar persons arising or incurred in the ordinary course of business);
17
(vi) create or hold capital stock in, any subsidiary that is not wholly owned (either directly or through one or more other subsidiaries) by this corporation, or permit any subsidiary to create, or authorize the creation of, or issue or obligate itself to issue, any shares of any class or series of capital stock, or sell, transfer or otherwise dispose of any capital stock of any direct or indirect subsidiary of this corporation, or permit any direct or indirect subsidiary to sell, lease, transfer, exclusively license or otherwise dispose (in a single transaction or series of related transactions) of all or substantially all of the assets of such subsidiary;
(vii) authorize, create or issue any equity security (including, without limitation, (A) any other security convertible into or exercisable for any such equity security or (B) any unit of debt and equity securities) having a preference over, or being on a parity with, any series of Preferred Stock with respect to dividends, liquidation or redemption, other than the issuance of any authorized but unissued shares of Preferred Stock designated in this Restated Certificate of Incorporation (including, without limitation, any security convertible into or exercisable for such shares of Preferred Stock);
(viii) (A) reclassify, alter or amend any existing security of this corporation that is pari passu with the Preferred Stock with respect to dividends, liquidation or redemption, if such reclassification, alteration or amendment would render such other security senior to the Preferred Stock in respect of any such right, preference or privilege or (B) reclassify, alter or amend any existing security of this corporation that is junior to the Preferred Stock with respect to dividends, liquidation or redemption, if such reclassification, alteration or amendment would render such other security senior to or pari passu with the Preferred Stock in respect of any such right, preference or privilege;
(ix) other than a redemption pursuant to Section 2(e), redeem, purchase or otherwise acquire (or pay into or set aside for a sinking fund for such purpose) any share or shares of Preferred Stock or Common Stock; provided, however, that this restriction shall not apply to the repurchase of shares of Common Stock from employees, officers, directors, consultants or other persons performing services for this corporation or any subsidiary pursuant to agreements approved by the Board of Directors under which this corporation has the option to repurchase such shares at the original per share purchase price (as adjusted for any stock splits, stock dividends, combinations, subdivisions, recapitalizations or the like) upon the occurrence of certain events, such as the termination of employment or service, or pursuant to a right of first refusal;
(x) redomicile this corporation to a jurisdiction other than the State of Delaware;
(xi) change the authorized number of, or method of electing the, directors of this corporation, or change the number of votes entitled to be cast by any director(s) on any matter; or
(xii) pay or declare any dividend or other distribution on any shares of capital stock of this corporation.
18
(b) Series C Preferred Stock Protective Provision. So long as 25% of the shares of Series C Preferred Stock originally issued (as adjusted for any stock splits, stock dividends, combinations, subdivisions, recapitalizations or the like with respect to such Preferred Stock) remains outstanding, this corporation shall not, either directly or indirectly by amendment, merger, domestication, transfer, continuance, reorganization, recapitalization, reclassification, waiver, statutory conversion, consolidation or otherwise) increase or decrease (other than by redemption or conversion) the total number of authorized shares of Series C Preferred Stock without (in addition to any other vote required by law or this Restated Certificate of Incorporation) obtaining the approval by vote or written consent, as provided by law, of the holders of sixty percent (60%) of the shares of the Series C Preferred Stock then outstanding (voting together as a single class and on an as-converted basis), and the taking of such act without the prior consent or vote being effected shall be null and void ab initio, and of no force or effect.
7. Status of Converted Stock. In the event any shares of Preferred Stock shall be converted pursuant to Section 4 hereof, the shares so converted shall be cancelled and shall not be issuable by this corporation. The Restated Certificate of Incorporation of this corporation shall be appropriately amended to effect the corresponding reduction in this corporation’s authorized capital stock.
8. Notices. Any notice required by the provisions of this Article IV(B) to be given to the holders of shares of Preferred Stock shall be deemed given (a) five (5) days following its deposit in the United States mail, postage prepaid, and addressed to each holder of record at his, her or its address appearing on the books of this corporation, (b) upon such notice being provided by electronic transmission in a manner permitted by the General Corporation Law or (c) five (5) days following such notice being provided in another manner then permitted by the General Corporation Law.
9. Waiver. Except as otherwise set forth herein, (a) subject to clause (b) below, any of the rights, powers, preferences and other terms of the Preferred Stock set forth herein may be waived on behalf of all holders of Preferred Stock by the affirmative written consent or vote of the holders that would otherwise be required to amend such right, powers, preferences, and other terms and (b) at any time more than one series of Preferred Stock is issued and outstanding, any of the rights, powers, preferences and other terms of any series of Preferred Stock set forth herein may be waived on behalf of all holders of such series of Preferred Stock only by the affirmative written consent or vote of the holders of such series that would otherwise be required to amend such right, power, preference, or other term.
C. Common Stock. The rights, preferences, privileges and restrictions granted to and imposed on the Common Stock are as set forth below in this Article IV(C).
1. Dividend Rights. Subject to the prior rights of holders of all classes of stock at the time outstanding having prior rights as to dividends, the holders of the Common Stock shall be entitled to receive, when, as and if declared by the Board of Directors, out of any assets of this corporation legally available therefor, any dividends as may be declared from time to time by the Board of Directors.
2. Liquidation Rights. Upon the liquidation, dissolution or winding up of this corporation, the assets of this corporation shall be distributed as provided in Section 2 of Article IV(B) hereof.
19
3. Redemption. The Common Stock is not redeemable at the option of the holder.
4. Voting Rights. The holder of each share of Common Stock shall have the right to one vote for each such share, and shall be entitled to notice of any stockholders’ meeting in accordance with the Bylaws of this corporation, and shall be entitled to vote upon such matters and in such manner as may be provided by law. The number of authorized shares of Common Stock may be increased or decreased (but not below the number of shares thereof then outstanding) by the affirmative vote of the holders of a majority of the stock of this corporation entitled to vote, irrespective of the provisions of Section 242(b)(2) of the General Corporation Law.
ARTICLE V
Except as otherwise provided in this Restated Certificate of Incorporation, in furtherance and not in limitation of the powers conferred by statute, the Board of Directors is expressly authorized to make, repeal, alter, amend and rescind any or all of the Bylaws of this corporation.
ARTICLE VI
Subject to the requirements of Section 6 of Article IV(B) hereof, the number of directors of this corporation shall be determined in the manner set forth in the Bylaws of this corporation. Unless otherwise provided herein, each director shall be entitled to one vote on each matter presented to the Board of Directors; provided that to the extent the approval of any particular director or directors is required by any agreement for specified actions, receipt of such approval shall be necessary for the Board of Directors to authorize such actions; provided further that, so long as the holders of Preferred Stock are entitled to elect a Preferred Director, the Requisite Preferred Director Vote shall be required for the authorization by the Board of Directors of any of the matters set forth in Section 3.12 of the Amended and Restated Investors’ Rights Agreement, dated as of or around the Series C Original Issue Date, by and among the corporation and the other parties thereto, as such agreement may be amended from time to time, to the extent required by such agreement and if any Preferred Director is then serving.
ARTICLE VII
Elections of directors need not be by written ballot unless the Bylaws of this corporation shall so provide.
ARTICLE VIII
Meetings of stockholders may be held within or without the State of Delaware, as the Bylaws of this corporation may provide. The books of this corporation may be kept (subject to any provision contained in the statutes) outside the State of Delaware at such place or places as may be designated from time to time by the Board of Directors or in the Bylaws of this corporation.
20
ARTICLE IX
To the fullest extent permitted by law, a director of this corporation shall not be personally liable to this corporation or its stockholders for monetary damages for breach of fiduciary duty as a director. If the General Corporation Law is amended after approval by the stockholders of this Article IX to authorize corporate action further eliminating or limiting the personal liability of directors, then the liability of a director of this corporation shall be eliminated or limited to the fullest extent permitted by the General Corporation Law as so amended.
Any amendment, repeal or modification of the foregoing provisions of this Article IX by the stockholders of this corporation shall not adversely affect any right or protection of a director of this corporation existing at the time of, or increase the liability of any director of this corporation with respect to any acts or omissions of such director occurring prior to, such amendment, repeal or modification.
ARTICLE X
This corporation reserves the right to amend, alter, change or repeal any provision contained in this Restated Certificate of Incorporation, in the manner now or hereafter prescribed by statute, and, subject to the requirements of the General Corporation Law and Section 6 of Article IV(B) hereof, all rights conferred upon stockholders herein are granted subject to this reservation.
ARTICLE XI
To the fullest extent permitted by applicable law, this corporation is authorized to provide indemnification of (and advancement of expenses to) directors, officers, employees and agents of this corporation (and any other persons to which General Corporation Law permits this corporation to provide indemnification) through Bylaw provisions, agreements with such persons, vote of stockholders or disinterested directors or otherwise, in excess of the indemnification and advancement otherwise permitted by Section 145 of the General Corporation Law.
Any amendment, repeal or modification of the foregoing provisions of this Article XI shall not adversely affect any right or protection of a director, officer, employee, agent or other person existing at the time of, or increase the liability of any such person with respect to any acts or omissions of such person occurring prior to, such amendment, repeal or modification.
ARTICLE XII
This corporation renounces any interest or expectancy of this corporation in, or in being offered an opportunity to participate in, any Excluded Opportunity. An “Excluded Opportunity” is any matter, transaction or interest that is presented to, or acquired, created or developed by, or which otherwise comes into the possession of, (i) any director of this corporation who is not an employee of this corporation or any of its subsidiaries, or (ii) any holder of Preferred Stock or any partner, member, director, stockholder, employee, affiliate or agent of any such holder, other than someone who is an employee of this corporation or any of its subsidiaries (collectively, “Covered Persons”), unless such matter, transaction or interest is presented to, or acquired, created or developed by, or otherwise comes into the possession of, a Covered Person expressly and solely in such Covered Person’s capacity as a director of this corporation while such
21
Covered Person is performing services in such capacity. Any repeal or modification of this Article XII will only be prospective and will not affect the rights under this Article XII in effect at the time of the occurrence of any actions or omissions to act giving rise to liability. Notwithstanding anything to the contrary contained elsewhere in this Restated Certificate of Incorporation, in addition to any other vote required by law or this Restated Certificate of Incorporation, the affirmative vote of the Requisite Holders will be required to amend or repeal, or to adopt any provisions inconsistent with this Article XII.
ARTICLE XIII
A. Forum Selection. Unless this corporation consents in writing to the selection of an alternative forum, the Court of Chancery of the State of Delaware (or, if the Court of Chancery does not have jurisdiction, the federal district court for the District of Delaware) shall, to the fullest extent permitted by law, be the sole and exclusive forum for (i) any derivative action or proceeding brought on behalf of this corporation, (ii) any action asserting a claim of breach of a fiduciary duty owed by any director, officer or other employee of this corporation to this corporation or this corporation’s stockholders, (iii) any action arising pursuant to any provision of the General Corporation Law or this Restated Certificate of Incorporation or the Bylaws (as either may be amended from time to time), or (iv) any action asserting a claim governed by the internal affairs doctrine, except for, as to each of (i) through (iv) above, any claim as to which the Court of Chancery determines that there is an indispensable party not subject to the jurisdiction of the Court of Chancery (and the indispensable party does not consent to the personal jurisdiction of the Court of Chancery within ten (10) days following such determination), which is vested in the exclusive jurisdiction of a court or forum other than the Court of Chancery, or for which the Court of Chancery does not have subject matter jurisdiction. Any person or entity purchasing or otherwise acquiring or holding any interest in shares of capital stock of this corporation shall be deemed to have notice of and consented to the provisions of this Article XIII.
B. Personal Jurisdiction. If any action the subject matter of which is within the scope of Article XIII(A) is filed in a court other than a court located within the State of Delaware (a “Foreign Action”) in the name of any stockholder, such stockholder shall be deemed to have consented to (i) the personal jurisdiction of the state and federal courts located within the State of Delaware in connection with any action brought in any such court to enforce Article XIII(A) (an “FSC Enforcement Action”) and (ii) having service of process made upon such stockholder in any such FSC Enforcement Action by service upon such stockholder’s counsel in the Foreign Action as agent for such stockholder.
ARTICLE XIV
Savings. If any provision or provisions of this Restated Certificate of Incorporation shall be held to be invalid, illegal or unenforceable as applied to any person or entity or circumstance for any reason whatsoever, then, to the fullest extent permitted by law, the validity, legality and enforceability of such provisions in any other circumstance and of the remaining provisions of this Restated Certificate of Incorporation (including, without limitation, each portion of any sentence of this Restated Certificate of Incorporation containing any such provision held to be invalid, illegal or unenforceable that is not itself held to be invalid, illegal or unenforceable) and the application of such provision to other persons or entities and circumstances shall not in any way be affected or impaired thereby.
* * *
22
THIRD: The foregoing amendment and restatement was approved by the holders of the requisite number of shares of said corporation in accordance with Section 228 of the General Corporation Law.
FOURTH: That said Restated Certificate of Incorporation, which restates and integrates and further amends the provisions of this corporation’s Restated Certificate of Incorporation, has been duly adopted in accordance with Sections 242 and 245 of the General Corporation Law.
23
IN WITNESS WHEREOF, this Restated Certificate of Incorporation has been executed by a duly authorized officer of this corporation on this 14th day of October, 2025.
| /s/ Quehuong (Kathy) Dong |
| Quehuong (Kathy) Dong |
| President and Chief Executive Officer |
CERTIFICATE OF AMENDMENT TO
THE AMENDED AND RESTATED
CERTIFICATE OF INCORPORATION OF
ELECTRA THERAPEUTICS, INC.
ELECTRA THERAPEUTICS, INC. (the “Company”), a corporation organized and existing under and by virtue of the provisions of the General Corporation Law of the State of Delaware (the “General Corporation Law”),
DOES HEREBY CERTIFY:
1. That the name of this corporation is Electra Therapeutics, Inc. and that this corporation was originally incorporated pursuant to the General Corporation Law on October 11, 2018.
2. That the Board of Directors of the Company duly adopted resolutions proposing to amend the current Amended and Restated Certificate of Incorporation of the Company (the “Restated Certificate”), declaring said amendment to be advisable and in the best interests of the Company and its stockholders, and authorizing the appropriate officers of the Company to solicit the consent of the stockholders therefor, which resolution setting forth the proposed amendment is as follows:
RESOLVED, that the Restated Certificate of the Company be amended by inserting the following into Article IV, Section A immediately before the first sentence:
“Effective upon the filing of this Certificate of Amendment to the Amended and Restated Certificate of Incorporation, with the Secretary of State of the State of Delaware (the “Effective Time”), every 1.5002 shares of Common Stock (as defined below) or Preferred Stock (as defined below) then issued and outstanding or held in the treasury of the corporation immediately prior to the Effective Time shall automatically be combined into one (1) share of Common Stock or Preferred Stock, as applicable, without any further action by the holders of such shares (the “Reverse Stock Split”). The Reverse Stock Split will be effected on a certificate-by-certificate basis, and any fractional shares resulting from such combination shall be rounded down to the nearest whole share on a certificate-by-certificate basis. No fractional shares shall be issued in connection with the Reverse Stock Split. In lieu of any fractional shares to which a holder would otherwise be entitled, the corporation shall pay cash equal to such fraction multiplied by the fair market value of a share of Common Stock as determined in good faith by the Board of Directors (as defined below). The Reverse Stock Split shall occur automatically without any further action by the holders of the shares of Common Stock and Preferred Stock affected thereby. All rights, preferences and privileges of the Common Stock and the Preferred Stock shall be appropriately adjusted to reflect the Reverse Stock Split in accordance with this Restated Certificate of Incorporation (as defined below).”
3. The foregoing amendment to the Restated Certificate has been approved by the holders of the requisite number of shares of the Company in accordance with Section 228 of the General Corporation Law.
4. The foregoing amendment to the Restated Certificate has been duly adopted in accordance with the provisions of Section 242 of the General Corporation Law.
* * * *
IN WITNESS WHEREOF, this Certificate of Amendment to the Amended and Restated Certificate of Incorporation has been executed by a duly authorized officer of this Company on this 10th day of September, 2026.
| ELECTRA THERAPEUTICS, INC. | ||
| By: | /s/ Quehuong (Kathy) Dong, Pharm.D., M.B.A. | |
| Quehuong (Kathy) Dong, Pharm.D., M.B.A. | ||
| President and Chief Executive Officer | ||
[Signature Page to Certificate of Amendment]
Exhibit 4.1
THIS CERTIFIES THAT is the owner of CUSIP DATED COUNTERSIGNED AND REGISTERED: COMPUTERSHARE TRUST COMPANY, N.A. TRANSFER AGENT AND REGISTRAR, FULLY-PAID AND NON-ASSESSABLE SHARES OF COMMON STOCK OF Electra Therapeutics, Inc. (hereinafter called the “Company”), transferable on the books of the Company in person or by duly authorized attorney, upon surrender of this Certificate properly endorsed. This Certificate and the shares represented hereby, are issued and shall be held subject to all of the provisions of the Certificate of Incorporation, as amended, and the By-Laws, as amended, of the Company (copies of which are on file with the Company and with the Transfer Agent), to all of which each holder, by acceptance hereof, assents. This Certificate is not valid unless countersigned and registered by the Transfer Agent and Registrar. Witness the facsimile seal of the Company and the facsimile signatures of its duly authorized officers. COMMON STOCK PAR VALUE $0.0001 COMMON STOCK SEE REVERSE FOR CERTAIN DEFINITIONS Certificate Number Shares . Electra Therapeutics, Inc. INCORPORATED UNDER THE LAWS OF THE STATE OF DELAWARE FACSIMILE SIGNATURE TO COME FACSIMILE SIGNATURE TO COME President Secretary By AUTHORIZED SIGNATURE ELECTRA THERAPEUTICS, INC. DELAWARE 10/11/2018 CORPORATE 284825 10 6 DD-MMM-YYYY **000000****************** ***000000***************** ****000000**************** *****000000*************** ******000000************** ** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Alexander David Sample **** Mr. Sample **** Mr. Sample **000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Sha res****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000 **Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****0 00000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares ****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**Shares****000000**S hares****000000**Shares****000000**Shares****000000**Shares**** ***ZERO HUNDRED THOUSAND ZERO HUNDRED AND ZERO*** MR. SAMPLE & MRS. SAMPLE & MR. SAMPLE & MRS. SAMPLE ZQ|CERT#|COY|CLS|RGSTRY|ACCT#|TRANSTYPE|RUN#|TRANS# ZQ00000000 Certificate Numbers 1234567890/1234567890 1234567890/1234567890 1234567890/1234567890 1234567890/1234567890 1234567890/1234567890 1234567890/1234567890 Total Transaction Num/No. 1 2 3 4 5 6 Denom. 1 2 3 4 5 6 Total 1 2 3 4 5 6 7 MR A SAMPLE DESIGNATION (IF ANY) ADD 1 ADD 2 ADD 3 ADD 4 PO Box 43004, Providence RI 02940-3004 CUSIP/IDENTIFIER XXXXXX XX X Holder ID XXXXXXXXXX Insurance Value 1,000,000.00 Number of Shares 123456 DTC 12345678 123456789012345 THIS CERTIFICATE IS TRANSFERABLE IN CITIES DESIGNATED BY THE TRANSFER AGENT, AVAILABLE ONLINE AT www.computershare.com electra therapeutics 1234567
The IRS requires that the named transfer agent (“we”) report the cost basis of certain shares or units acquired after January 1, 2011. If your shares or units are covered by the legislation, and you requested to sell or transfer the shares or units using a specific cost basis calculation method, then we have processed as you requested. If you did not specify a cost basis calculation method, then we have defaulted to the first in, first out (FIFO) method. Please consult your tax advisor if you need additional information about cost basis. If you do not keep in contact with the issuer or do not have any activity in your account for the time period specified by state law, your property may become subject to state unclaimed property laws and transferred to the appropriate state. . ELECTRA THERAPEUTICS, INC. THE COMPANY WILL FURNISH WITHOUT CHARGE TO EACH STOCKHOLDER WHO SO REQUESTS, A SUMMARY OF THE POWERS, DESIGNATIONS, PREFERENCES AND RELATIVE, PARTICIPATING, OPTIONAL OR OTHER SPECIAL RIGHTS OF EACH CLASS OF STOCK OF THE COMPANY AND THE QUALIFICATIONS, LIMITATIONS OR RESTRICTIONS OF SUCH PREFERENCES AND RIGHTS, AND THE VARIATIONS IN RIGHTS, PREFERENCES AND LIMITATIONS DETERMINED FOR EACH SERIES, WHICH ARE FIXED BY THE CERTIFICATE OF INCORPORATION OF THE COMPANY, AS AMENDED, AND THE RESOLUTIONS OF THE BOARD OF DIRECTORS OF THE COMPANY, AND THE AUTHORITY OF THE BOARD OF DIRECTORS TO DETERMINE VARIATIONS FOR FUTURE SERIES. SUCH REQUEST MAY BE MADE TO THE OFFICE OF THE SECRETARY OF THE COMPANY OR TO THE TRANSFER AGENT. THE BOARD OF DIRECTORS MAY REQUIRE THE OWNER OF A LOST OR DESTROYED STOCK CERTIFICATE, OR HIS LEGAL REPRESENTATIVES, TO GIVE THE COMPANY A BOND TO INDEMNIFY IT AND ITS TRANSFER AGENTS AND REGISTRARS AGAINST ANY CLAIM THAT MAY BE MADE AGAINST THEM ON ACCOUNT OF THE ALLEGED LOSS OR DESTRUCTION OF ANY SUCH CERTIFICATE. For US purposes the following abbreviations, when used in the inscription on the face of this certificate, shall be construed as though they were written out in full according to applicable laws or regulations: TEN COM - as tenants in common UNIF GIFT MIN ACT -.............................................Custodian................................................. ( Cust) (Minor) TEN ENT - as tenants by the entireties under Uniform Gifts to Minors Act......................................................... (State) JT TEN - as joint tenants with right of survivorship UNIF TRF MIN ACT -.............................................Custodian (until age.................................) and not as tenants in common (Cust) ..............................under Uniform Transfers to Minors Act.................... (Minor) (State) Additional abbreviations may also be used though not in the above list. For value received, hereby sell, assign and transfer unto Shares Attorney Dated: 20 Signature: Signature: Signature(s) Guaranteed: Medallion Guarantee Stamp THE SIGNATURE(S) SHOULD BE GUARANTEED BY AN ELIGIBLE GUARANTOR INSTITUTION (Banks, Stockbrokers, Savings and Loan Associations and Credit Unions) WITH MEMBERSHIP IN AN APPROVED SIGNATURE GUARANTEE MEDALLION PROGRAM, PURSUANT TO S.E.C. RULE 17Ad-15. PLEASE INSERT SOCIAL SECURITY OR OTHER IDENTIFYING NUMBER OF ASSIGNEE (PLEASE PRINT OR TYPEWRITE NAME AND ADDRESS, INCLUDING POSTAL ZIP CODE, OF ASSIGNEE) of the common stock represented by the within Certificate, and do hereby irrevocably constitute and appoint to transfer the said stock on the books of the within-named Company with full power of substitution in the premises. Notice: The signature to this assignment must correspond with the name as written upon the face of the certificate, in every particular, without alteration or enlargement, or any change whatever. 1234567 security I instruct ions this watermarked paper, do not accept with out noting watermark hold to light to verify watermark.
| Charles J. Bair T: +1 858 550 6142 cbair@cooley.com |
Exhibit 5.1 |
September 14, 2026
Electra Therapeutics, Inc.
230 E Grand Avenue, Suite S-100
South San Francisco, California 94080
Ladies and Gentlemen:
We have acted as counsel to Electra Therapeutics, Inc., a Delaware corporation (the “Company”), in connection with the filing by the Company of a registration statement (No. 333-298617) on Form S-1 (the “Registration Statement”) with the Securities and Exchange Commission (the “Commission”), including a related prospectus included in the Registration Statement (the “Prospectus”), covering an underwritten public offering by the Company of up to 24,916,667 shares (the “Shares”) of its common stock, par value $0.0001 per share, including up to 3,250,000 Shares that may be sold pursuant to the exercise of an option to purchase additional Shares granted to the underwriters.
In connection with this opinion, we have (i) examined and relied upon (a) the Registration Statement and the Prospectus, (b) the Company’s certificate of incorporation and bylaws, each as currently in effect, (c) the Company’s Amended and Restated Certificate of Incorporation and Amended and Restated Bylaws, filed as Exhibits 3.2 and 3.4 to the Registration Statement, respectively, each of which is to be in effect in connection with the closing of the offering contemplated by the Registration Statement and (d) such other records, documents, opinions, certificates, memoranda and instruments as in our judgment are necessary or appropriate to enable us to render the opinion expressed below and (ii) assumed that (a) the Shares will be sold at a price established by the Board of Directors of the Company or a duly authorized committee thereof and (b) the Amended and Restated Certificate of Incorporation referred to in clause (i)(c) is filed with the Secretary of State of the State of Delaware before issuance of the Shares.
We have assumed the genuineness of all signatures, the authenticity of all documents submitted to us as originals, the conformity to originals of all documents submitted to us as copies, the accuracy, completeness and authenticity of certificates of public officials and the due authorization, execution and delivery of all documents by all persons other than the Company. As to certain factual matters, we have relied upon a certificate of an officer of the Company and have not independently verified such matters.
Our opinion is expressed only with respect to the General Corporation Law of the State of Delaware. We express no opinion to the extent that any other laws are applicable to the subject matter hereof and express no opinion and provide no assurance as to compliance with any federal or state securities law, rule or regulation.
On the basis of the foregoing, in reliance thereon and subject to the assumptions, qualifications, limitations and exceptions set forth herein, we are of the opinion that the Shares, when sold and issued against payment therefor as described in the Registration Statement and the Prospectus, will be validly issued, fully paid and nonassessable.
Cooley LLP 10265 Science Center Drive, San Diego, CA 92121-1117
t: +1 (858) 550-6000 f: +1 (858) 550-6420 cooley.com
Electra Therapeutics, Inc.
Page 2
This opinion is limited to the matters expressly set forth in this letter, and no opinion has been or should be implied, or may be inferred, beyond the matters expressly stated. This opinion speaks only as to law and facts in effect or existing as of the date hereof, and we have no obligation or responsibility to update or supplement this letter to reflect any facts or circumstances that may hereafter come to our attention or any changes in law that may hereafter occur.
We consent to the reference to our firm under the heading “Legal Matters” in the Prospectus and to the filing of this opinion as an exhibit to the Registration Statement. In giving such consents, we do not thereby admit that we are in the category of persons whose consent is required under Section 7 of the Securities Act, or the rules and regulations of the Commission thereunder.
Sincerely,
Cooley LLP
| By: | /s/ Charles J. Bair | |
| Charles J. Bair |
Cooley LLP 10265 Science Center Dr., San Diego, CA 92121
t: +1 (858) 550-6000 f: +1 (858) 550-6420 cooley.com
Exhibit 10.2
ELECTRA THERAPEUTICS, INC.
2026 EQUITY INCENTIVE PLAN
ADOPTED BY THE BOARD OF DIRECTORS: AUGUST 19, 2026
APPROVED BY THE STOCKHOLDERS: SEPTEMBER 10, 2026
| 1. | GENERAL. |
(a) Plan Purpose. The Company, by means of the Plan, seeks to secure and retain the services of Employees, Directors and Consultants, to provide incentives for such persons to exert maximum efforts for the success of the Company and any Affiliate and to provide a means by which such persons may be given an opportunity to benefit from increases in value of the Common Stock through the granting of Awards.
(b) Types of Awards. The Plan provides for the grant of the following Awards: (i) Incentive Stock Options; (ii) Nonstatutory Stock Options; (iii) Stock Appreciation Rights; (iv) Restricted Stock Awards; (v) Restricted Stock Unit Awards; (vi) Performance Awards; and (vii) Other Awards.
(c) Effective Date. The Plan will come into existence on the Effective Date and no Award may be granted prior to the Effective Date.
2. SHARES SUBJECT TO THE PLAN.
(a) Share Reserve. Subject to adjustment in accordance with Section 2(c) and any adjustments as necessary to implement any Capitalization Adjustments, the aggregate number of Shares that may be issued pursuant to Awards will not exceed 13,446,281, which is the sum of: (i) 7,697,039 Shares, plus (ii) up to 5,749,242 Returning Shares, as they become available from time to time. In addition, subject to any adjustments as necessary to implement any Capitalization Adjustments, the aggregate number of Shares will automatically increase on January 1 of each year for a period of ten years commencing January 1, 2027 and ending on (and including) January 1, 2036, in an amount equal to 5% of the total number of shares of Fully Diluted Capital Stock outstanding on December 31 of the preceding year; provided, however, that the Board may act prior to January 1st of any given year to provide that the increase for that year will be a lesser number of Shares.
(b) Aggregate Incentive Stock Option Limit. The aggregate maximum number of Shares that may be issued pursuant to the exercise of Incentive Stock Options is specified in Section 3(b)(iv).
(c) Share Reserve Operation.
(i) Limit Applies to Common Stock Issued Pursuant to Awards. For clarity, the Share Reserve is a limit on the number of shares of Common Stock that may be issued pursuant to Awards and does not limit the granting of Awards, except that the Company will keep available at all times the number of shares of Common Stock reasonably required to satisfy its obligations to issue shares pursuant to such Awards. Shares may be issued in connection with a merger or acquisition as permitted by, as applicable, Nasdaq Listing Rule 5635(c)(3), NYSE Listed Company Manual Section 303A.08, NYSE American Company Guide Section 711 or other applicable rule, and such issuance will not reduce the number of shares available for issuance under the Plan.
1
(ii) Actions that Do Not Constitute Issuance of Shares and Do Not Reduce Share Reserve. The following actions do not result in an issuance of Shares under the Plan and accordingly do not reduce the number of Shares subject to the Share Reserve and available for issuance under the Plan: (1) the expiration or termination of any portion of an Award without the Shares covered by that portion of the Award having been issued; (2) the settlement of any portion of an Award in cash rather than Shares; (3) the withholding of Shares that would otherwise be issued by the Company to satisfy the exercise, strike or purchase price of an Award; or (4) the withholding of Shares that would otherwise be issued by the Company to satisfy a tax withholding obligation in connection with an Award.
(iii) Reversion of Previously Issued Shares to Share Reserve. The following Shares previously issued pursuant to an Award and accordingly initially deducted from the Share Reserve will be added back to the Share Reserve and again become available for issuance under the Plan: (1) any Shares that are forfeited back to or repurchased by the Company because of a failure to meet a contingency or condition required for the vesting of the Award; (2) any Shares that are reacquired by the Company to satisfy the exercise, strike or purchase price of an Award; and (3) any Shares that are reacquired by the Company to satisfy a tax withholding obligation in connection with an Award.
3. ELIGIBILITY AND LIMITATIONS.
(a) Eligible Award Recipients. Subject to the terms of the Plan, Employees, Directors, and Consultants are eligible to receive Awards.
(b) Specific Award Limitations.
(i) Limitations on Incentive Stock Option Recipients. Incentive Stock Options may be granted only to Employees of the Company or a “parent corporation” or “subsidiary corporation” thereof (as such terms are defined in Sections 424(e) and (f) of the Code).
(ii) Incentive Stock Option $100,000 Limitation. To the extent that the aggregate Fair Market Value (determined at the time of grant) of Shares with respect to which Incentive Stock Options are exercisable for the first time by any Participant during any calendar year (under all plans of the Company and any “parent corporation” or “subsidiary corporation” thereof (as such terms are defined in Sections 424(e) and (f) of the Code)) exceeds $100,000 (or such other limit established in the Code) or otherwise does not comply with the rules governing Incentive Stock Options, the Options or portions thereof that exceed such limit (according to the order in which they were granted) or otherwise do not comply with such rules will be treated as Nonstatutory Stock Options, notwithstanding any contrary provision of the applicable Award Agreement(s).
(iii) Limitations on Incentive Stock Options Granted to Ten Percent Stockholders. A Ten Percent Stockholder may not be granted an Incentive Stock Option unless (1) the per Share exercise price of such Option is at least 110% of the Fair Market Value of a Share on the date of grant of such Option and (2) the Option is not exercisable after the expiration of five years from the date of grant of such Option.
(iv) Aggregate Incentive Stock Option Limit. Notwithstanding anything to the contrary in Section 2(a) and subject to any adjustments as necessary to implement any Capitalization Adjustments, the aggregate maximum number of Shares that may be issued pursuant to the exercise of Incentive Stock Options is 40,338,843 Shares.
(c) Non-Employee Director Compensation Limit. The aggregate value of all compensation granted or paid, as applicable, to any individual for service as a Non-Employee Director with respect to any calendar year, including Awards granted and cash fees paid by the Company to such Non-Employee Director, will not exceed (1) $750,000 in total value or (2) in the event such Non-Employee Director is first appointed or elected to the Board during such calendar year, $1,000,000 in total value, in each case, calculating the value of any equity awards based on the grant date fair value of such equity awards for financial reporting purposes. The limitations in this Section 3(c) shall apply commencing with the first calendar year that begins following the Effective Date.
2
4. OPTIONS AND STOCK APPRECIATION RIGHTS.
Each Option and Stock Appreciation Right will have such terms and conditions as determined by the Board. Each Option will be designated in writing as an Incentive Stock Option or Nonstatutory Stock Option at the time of grant; provided, however, that if an Option is not so designated or if an Option designated as an Incentive Stock Option fails to qualify as an Incentive Stock Option, then such Option will be a Nonstatutory Stock Option, and the Shares purchased upon exercise of each type of Option will be separately accounted for. Each Stock Appreciation Right will be denominated in Share equivalents. The terms and conditions of separate Options and Stock Appreciation Rights need not be identical; provided, however, that each Award Agreement will conform (through incorporation of provisions hereof by reference in the Award Agreement or otherwise) to the substance of each of the following provisions:
(a) Term. Subject to Section 3(b) regarding Ten Percent Stockholders, no Option or Stock Appreciation Right will be exercisable after the expiration of ten years from the date of grant of such Award or such shorter period specified in the Award Agreement.
(b) Exercise or Strike Price. Subject to Section 3(b) regarding Ten Percent Stockholders, the per Share exercise or strike price of each Option or Stock Appreciation Right will not be less than 100% of the Fair Market Value of a Share on the date of grant of such Award. Notwithstanding the foregoing, an Option or Stock Appreciation Right may be granted with a per Share exercise or strike price lower than 100% of the Fair Market Value of a Share on the date of grant of such Award if such Award is granted pursuant to an assumption of or substitution for another option or stock appreciation right pursuant to a Corporate Transaction and in a manner consistent with the provisions of Sections 409A and, if applicable, 424(a) of the Code.
(c) Exercise Procedure and Payment of Exercise Price for Options. In order to exercise an Option, the Participant must provide notice of exercise to the Plan Administrator in accordance with the procedures specified in the Award Agreement or otherwise provided by the Company. The Board has the authority to grant Options that do not permit all of the following methods of payment (or otherwise restrict the ability to use certain methods) and to grant Options that require the consent of the Company to utilize a particular method of payment. The exercise price of an Option may be paid, to the extent permitted by Applicable Law and as determined by the Board, by one or more of the following methods of payment to the extent set forth in the Award Agreement:
(i) cash or check, bank draft or money order payable to the Company;
(ii) consideration received by the Company under a broker-assisted (or other) cashless exercise program (whether through a broker or otherwise) implemented by the Company in connection with the Plan;
(iii) other Shares, provided that such Shares have a Fair Market Value on the date of surrender equal to the aggregate exercise price of the Shares as to which the Option will be exercised and provided further that accepting the Shares will not result in any adverse accounting consequences to the Company, as the Company determines in its sole discretion;
(iv) if the Option is a Nonstatutory Stock Option, a “net exercise” arrangement; or
3
(v) any other form of consideration that may be acceptable to the Board and permissible under Applicable Law.
(d) Exercise Procedure and Payment of Appreciation Distribution for Stock Appreciation Rights. In order to exercise any Stock Appreciation Right, the Participant must provide notice of exercise to the Plan Administrator in accordance with the Award Agreement. The appreciation distribution payable to a Participant upon the exercise of a Stock Appreciation Right will not be greater than an amount equal to the excess of (i) the aggregate Fair Market Value on the date of exercise of a number of Shares equal to the number of Share equivalents that are vested and being exercised under the Stock Appreciation Right, over (ii) the aggregate strike price of the portion of the Stock Appreciation Right that is being exercised. The appreciation distribution may be paid to the Participant in the form of Shares or cash (or any combination of Shares and cash), as determined by the Board and specified in the Award Agreement.
(e) Transferability. Options and Stock Appreciation Rights may not be transferred to third party financial institutions for value. The Board may impose such additional limitations on the transferability of an Option or Stock Appreciation Right as it determines. In the absence of any such determination by the Board, the following restrictions on the transferability of Options and Stock Appreciation Rights will apply, provided that except as explicitly provided herein, neither an Option nor a Stock Appreciation Right may be transferred for consideration and provided, further, that if an Option is an Incentive Stock Option, such Option may be deemed to be a Nonstatutory Stock Option as a result of such transfer:
(i) Restrictions on Transfer. An Option or Stock Appreciation Right will not be transferable, except by will or by the laws of descent and distribution, and will be exercisable during the lifetime of the Participant only by the Participant; provided, however, that the Board may permit transfer of an Option or Stock Appreciation Right in a manner that is not prohibited by applicable tax and securities laws upon the Participant’s request, including to a trust if the Participant is considered to be the sole beneficial owner of such trust (as determined under Section 671 of the Code and applicable state law) while such Option or Stock Appreciation Right is held in such trust, provided that the Participant and the trustee enter into a transfer and other agreements required by the Company.
(ii) Domestic Relations Orders. Notwithstanding the foregoing, subject to the execution of transfer documentation in a format acceptable to the Company and subject to the approval of the Board or a duly authorized Officer, an Option or Stock Appreciation Right may be transferred pursuant to a domestic relations order.
(f) Vesting. The Board may impose such restrictions on or conditions to the vesting and exercisability of an Option or Stock Appreciation Right as determined by the Board. Except as otherwise provided in the applicable Award Agreement or other written agreement between a Participant and the Company or an Affiliate that is approved by the Board, vesting of Options and Stock Appreciation Rights will cease upon termination of the Participant’s Continuous Service.
(g) Termination of Continuous Service for Cause. Except as explicitly otherwise provided in the Award Agreement or other written agreement between a Participant and the Company or an Affiliate that is approved by the Board, if a Participant’s Continuous Service is terminated for Cause, the Participant’s Options and Stock Appreciation Rights will terminate and be forfeited immediately upon such termination of Continuous Service, and the Participant will be prohibited from exercising any portion (including any vested portion) of such Awards on and after the date of such termination of Continuous Service and the Participant will have no further right, title or interest in such forfeited Award, the Shares subject to the forfeited Award, or any consideration in respect of the forfeited Award.
4
(h) Post-Termination Exercise Period Following Termination of Continuous Service for Reasons Other than Cause. Subject to Sections 4(i) and 6, if a Participant’s Continuous Service terminates for any reason other than for Cause, the Participant may exercise their Option or Stock Appreciation Right to the extent vested, but only within the following period of time or, if applicable, such other period of time provided in the Award Agreement or other written agreement between a Participant and the Company or an Affiliate that is approved by the Board; provided, however, that in no event may such Award be exercised after the expiration of its maximum term (as set forth in Section 4(a)):
(i) three months following the date of the termination if the termination is a termination without Cause (other than any termination due to the Participant’s Disability or death);
(ii) 12 months following the date of the termination if the termination is due to the Participant’s Disability;
(iii) 18 months following the date of the termination if the termination is due to the Participant’s death; or
(iv) 18 months following the date of the Participant’s death if the death occurs following the date of the termination but during the period the Award is otherwise exercisable (as provided in (i) or (ii) above).
Following the date of termination, to the extent the Participant does not exercise an Award within the applicable Post-Termination Exercise Period (or, if earlier, prior to the expiration of the maximum term of such Award), the unexercised portion of the Award will terminate, and the Participant will have no further right, title or interest in the terminated Award, the Shares subject to the terminated Award, or any consideration in respect of the terminated Award.
(i) Restrictions on Exercise; Extension of Exercisability. A Participant may not exercise an Option or Stock Appreciation Right at any time that the issuance of Shares upon exercise of the Award would violate Applicable Law. Except as otherwise provided in the Award Agreement or other written agreement between a Participant and the Company or an Affiliate that is approved by the Board, if a Participant’s Continuous Service terminates for any reason other than for Cause and, at any time during the last thirty days of the applicable Post-Termination Exercise Period: (i) the exercise of the Participant’s Option or Stock Appreciation Right would be prohibited solely because the issuance of Shares upon exercise of the Award would violate Applicable Law, or (ii) the immediate sale of any Shares issued upon the exercise of the Award would violate the Company’s Trading Policy, then the applicable Post-Termination Exercise Period will be extended to the last day of the calendar month that commences following the date the exercise period would otherwise expire, with an additional extension of the exercise period to the last day of the next calendar month to apply if any of the foregoing restrictions apply at any time during the extended exercise period, generally without limitation as to the maximum permitted number of extensions; provided, however, that in no event may such Award be exercised after the expiration of its maximum term (as set forth in Section 4(a)).
(j) Whole Shares. Options and Stock Appreciation Rights may be exercised only with respect to whole Shares or their equivalents.
| 5. | RESTRICTED STOCK AWARDS, RESTRICTED STOCK UNIT AWARDS, AND OTHER AWARDS. |
(a) Restricted Stock Awards and Restricted Stock Unit Awards. Each Restricted Stock Award and Restricted Stock Unit Award will have such terms and conditions as determined by the Board; provided, however, that each Award Agreement will conform (through incorporation of the provisions hereof by reference in the Award Agreement or otherwise) to the substance of each of the following provisions:
5
(i) Form of Award.
(1) Restricted Stock Awards: At the Board’s election, Shares subject to a Restricted Stock Award may be (A) held in book entry form subject to the Company’s instructions until the Shares become vested or any other restrictions lapse, or (B) evidenced by a certificate, which certificate will be held in the form and manner as determined by the Board. Unless otherwise determined by the Board, a Participant will have voting and other rights as a stockholder of the Company with respect to any Shares subject to a Restricted Stock Award.
(2) Restricted Stock Unit Awards: A Restricted Stock Unit Award represents a Participant’s right to be issued on a future date the number of Shares that is equal to the number of units subject to the Restricted Stock Unit Award. As a holder of a Restricted Stock Unit Award, a Participant is an unsecured creditor of the Company with respect to the Company’s unfunded obligation, if any, to issue Shares in settlement of the Award and nothing contained in the Plan or any Award Agreement, and no action taken pursuant to its provisions, will create or be construed to create a trust of any kind or a fiduciary relationship between a Participant and the Company or an Affiliate or any other person. A Participant will not have voting or any other rights as a stockholder of the Company with respect to any Restricted Stock Unit Award (unless and until Shares are actually issued in settlement of a vested Restricted Stock Unit Award).
(ii) Consideration.
(1) Restricted Stock Awards: A Restricted Stock Award may be granted in consideration for (A) cash or check, bank draft or money order payable to the Company, (B) services to the Company or an Affiliate, or (C) any other form of consideration as the Board may determine and permissible under Applicable Law.
(2) Restricted Stock Unit Awards: Unless otherwise determined by the Board at the time of grant, a Restricted Stock Unit Award will be granted in consideration for the Participant’s services to the Company or an Affiliate, such that the Participant will not be required to make any payment to the Company (other than such services) with respect to the grant or vesting of the Restricted Stock Unit Award, or the issuance of any Shares pursuant to the Restricted Stock Unit Award. If, at the time of grant, the Board determines that any consideration must be paid by the Participant (in a form other than the Participant’s services to the Company or an Affiliate) upon the issuance of any Shares in settlement of the Restricted Stock Unit Award, such consideration may be paid in any form of consideration as the Board may determine and that is permissible under Applicable Law.
(iii) Vesting. The Board may impose such restrictions on or conditions to the vesting of a Restricted Stock Award or Restricted Stock Unit Award as determined by the Board. Except as otherwise provided in the Award Agreement or other written agreement between a Participant and the Company or an Affiliate that is approved by the Board, vesting of Restricted Stock Awards and Restricted Stock Unit Awards will cease upon termination of the Participant’s Continuous Service.
(iv) Termination of Continuous Service. Except as otherwise provided in the Award Agreement or other written agreement between a Participant and the Company or an Affiliate that is approved by the Board, if a Participant’s Continuous Service terminates for any reason, (1) the Company may receive through a forfeiture condition or a repurchase right any or all of the Shares held by the Participant under their Restricted Stock Award that have not vested as of the date of such termination as set
6
forth in the Award Agreement and the Participant will have no further right, title or interest in the Restricted Stock Award, the Shares subject to the Restricted Stock Award, or any consideration in respect of the Restricted Stock Award and (2) any portion of the Participant’s Restricted Stock Unit Award that has not vested will be forfeited upon such termination and the Participant will have no further right, title or interest in the Restricted Stock Unit Award, the Shares issuable pursuant to the Restricted Stock Unit Award, or any consideration in respect of the Restricted Stock Unit Award.
(v) Dividends and Dividend Equivalents. Dividends or dividend equivalents may be paid or credited, as applicable, with respect to any Shares subject to a Restricted Stock Award or Restricted Stock Unit Award, as determined by the Board and specified in the Award Agreement.
(vi) Settlement of Restricted Stock Unit Awards. A Restricted Stock Unit Award may be settled by the issuance of Shares or cash (or any combination thereof), as determined by the Board and specified in the Award Agreement. At the time of grant, the Board may determine to impose such restrictions or conditions that delay such delivery to a date following the vesting of the Restricted Stock Unit Award.
(b) Performance Awards. With respect to any Performance Award, the length of any Performance Period, the Performance Goals to be achieved during the Performance Period, the other terms and conditions of such Award, and the measure of whether and to what degree such Performance Goals have been attained will be determined by the Board.
(c) Other Awards. Other forms of Awards valued in whole or in part by reference to, or otherwise based on, Shares, including the appreciation in value thereof may be granted either alone or in addition to Awards provided for under Section 4 and the preceding provisions of this Section 5. Subject to the provisions of the Plan, the Board will have sole and complete discretion to determine the persons to whom and the time or times at which such Other Awards will be granted, the number of Shares (or the cash equivalent thereof) to be granted pursuant to such Other Awards and all other terms and conditions of such Other Awards.
6. ADJUSTMENTS UPON CHANGES IN COMMON STOCK; OTHER CORPORATE EVENTS.
(a) Capitalization Adjustments. In the event of a Capitalization Adjustment, the Board shall appropriately and proportionately adjust: (i) the class(es) and maximum number of Shares subject to the Plan; (ii) the class(es) and maximum number of Shares that may be issued pursuant to the exercise of Incentive Stock Options pursuant to Section 3(b)(iv); and (iii) the class(es) and number of underlying Shares and the exercise price, strike price or purchase price of outstanding Awards. The Board shall make such adjustments, and its determination shall be final, binding, and conclusive. Notwithstanding the foregoing, no fractional Shares or rights shall be created in order to implement any Capitalization Adjustment. The Board shall determine an appropriate equivalent benefit, if any, for any fractional Shares or rights that might be created by the adjustments referred to in the preceding provisions of this Section.
(b) Dissolution or Liquidation. Except as otherwise provided in the Award Agreement, in the event of a dissolution or liquidation of the Company, all outstanding Awards (other than Awards consisting of vested and outstanding Shares not subject to a forfeiture condition or the Company’s right of repurchase) will terminate immediately prior to the completion of such dissolution or liquidation, and the Shares subject to the Company’s repurchase rights or subject to a forfeiture condition may be repurchased or reacquired by the Company notwithstanding the fact that the holder of such Award is providing Continuous Service, provided, however, that the Board may determine to cause some or all Awards to become fully vested, exercisable or no longer subject to repurchase or forfeiture (to the extent such Awards have not previously expired or terminated) before the dissolution or liquidation is completed but contingent on its completion.
7
(c) Corporate Transaction. The following provisions will apply to Awards in the event of a Corporate Transaction, unless otherwise provided in the instrument evidencing the Award or any other written agreement between the Company or any Affiliate and the Participant that is approved by the Board or unless otherwise expressly provided by the Board at the time of grant of an Award.
(i) Awards May Be Assumed. In the event of a Corporate Transaction, any surviving corporation or acquiring corporation (or the surviving or acquiring corporation’s parent company) may assume or continue any or all Awards outstanding under the Plan or may substitute similar awards for Awards outstanding under the Plan (including but not limited to, awards to acquire the same consideration paid to the stockholders of the Company pursuant to the Corporate Transaction), and any reacquisition or repurchase rights held by the Company in respect of Shares issued pursuant to Awards may be assigned by the Company to the successor of the Company (or the successor’s parent company, if any), in connection with such Corporate Transaction. A surviving corporation or acquiring corporation (or its parent) may choose to assume or continue only a portion of an Award or substitute a similar award for only a portion of an Award, or may choose to assume, continue or substitute the Awards held by some, but not all Participants. The terms of any assumption, continuation or substitution will be determined by the Board.
(ii) Awards Held by Current Participants. In the event of a Corporate Transaction in which the surviving corporation or acquiring corporation (or its parent company) does not assume or continue such outstanding Awards (or portions thereof) or substitute similar awards for such outstanding Awards (or portions thereof), then with respect to any Award (or portion thereof) that has not been assumed, continued or substituted and that is held by a Participant whose Continuous Service has not terminated prior to the effective time of the Corporate Transaction (referred to as the “Current Participants”), the vesting of such Award or portion thereof (and, with respect to an Option or Stock Appreciation Right, the time when such Award or portion thereof may be exercised) will be accelerated in full to a date prior to the effective time of such Corporate Transaction (contingent upon the effectiveness of the Corporate Transaction) as the Board determines (or, if the Board does not determine such a date, to the date that is five days prior to the effective time of the Corporate Transaction), and such Award (or portion thereof) will terminate if not exercised (if applicable) at or prior to the effective time of the Corporate Transaction, and any reacquisition or repurchase rights held by the Company with respect to such Award (or portion thereof) will lapse (contingent upon the effectiveness of the Corporate Transaction). With respect to the vesting of a Performance Award (or portion thereof) that will accelerate upon the occurrence of a Corporate Transaction pursuant to this subsection (ii), the vesting of such Performance Award (or portion thereof) will accelerate at 100% of the target level upon the occurrence of the Corporate Transaction in which the Award (or portion thereof) is not assumed, continued or substituted in accordance with Section 6(c)(i). With respect to the vesting of Awards (or portions thereof) that will accelerate upon the occurrence of a Corporate Transaction pursuant to this subsection (ii) and are settled in the form of a cash payment, such cash payment will be made no later than 30 days following the occurrence of the Corporate Transaction or such later date as required to comply with Section 409A of the Code.
(iii) Awards Held by Persons other than Current Participants. In the event of a Corporate Transaction in which the surviving corporation or acquiring corporation (or its parent company) does not assume or continue such outstanding Awards or substitute similar awards for such outstanding Awards, then with respect to Awards that have not been assumed, continued or substituted and that are held by persons other than Current Participants, the Awards will terminate if not exercised (if applicable) prior to the occurrence of the Corporate Transaction; provided, however, that any reacquisition or repurchase rights held by the Company with respect to such Awards will not terminate and may continue to be exercised notwithstanding the Corporate Transaction.
8
(iv) Payment for Awards. Notwithstanding the foregoing, in the event any portion of an Award will terminate prior to the effective time of a Corporate Transaction, the Board may provide, in its sole discretion, that the holder of the Award will receive a payment, in such form as may be determined by the Board, equal in value, at the effective time, to the excess, if any, of (1) the value of the property the Participant would have received with respect to that portion of the Award (including, at the discretion of the Board, any unvested portion of such Award), over (2) the exercise price payable by such holder in connection with that portion of the Award, if any.
(d) Appointment of Stockholder Representative. As a condition to the receipt of an Award under this Plan, a Participant will be deemed to have agreed that the Award will be subject to the terms of any agreement governing a Corporate Transaction involving the Company, including, without limitation, a provision for the appointment of a stockholder representative that is authorized to act on the Participant’s behalf with respect to any escrow, indemnities and any contingent consideration.
(e) No Restriction on Right to Undertake Transactions. The grant of any Award under the Plan and the issuance of Shares pursuant to any Award does not affect or restrict in any way the right or power of the Company, the Board or the stockholders of the Company to make or authorize any adjustment, recapitalization, reorganization or other change in the Company’s capital structure or its business, any Change in Control, any Corporate Transaction, any merger or consolidation of the Company, any issue of stock or of options, rights or options to purchase stock or of bonds, debentures, preferred or prior preference stocks whose rights are superior to or affect the Common Stock or the rights thereof or which are convertible into or exchangeable for Common Stock, or the dissolution or liquidation of the Company, or any sale or transfer of all or any part of its assets or business, or any other corporate act or proceeding, whether of a similar character or otherwise.
7. ADMINISTRATION.
(a) Administration by Board. The Board will administer the Plan unless and until the Board delegates administration of the Plan to a Committee or Committees, as provided in subsection (c) below.
(b) Powers of Board. The Board will have the power, subject to, and within the limitations of, the express provisions of the Plan:
(i) To determine from time to time (1) which of the persons eligible under the Plan will be granted Awards; (2) when and how each Award will be granted; (3) what type or combination of types of Award will be granted; (4) the terms and conditions of each Award granted (which need not be identical), including the time or times when a person will be permitted to receive an issuance of Shares or other payment pursuant to an Award; (5) the number of Shares or cash equivalent with respect to which an Award will be granted to each such person; (6) the value applicable to an Award; and (7) the terms of any Performance Award that is not valued in whole or in part by reference to, or otherwise based on, Shares, including the amount of cash payment that may be earned and the timing of payment.
(ii) To construe and interpret the Plan and Awards granted under it, and to establish, amend and revoke rules and regulations for its administration. The Board, in the exercise of this power, may correct any defect, omission or inconsistency in the Plan, any Award Agreement, or any written agreement between the Company or any Affiliate and a Participant that is approved by the Board (to the extent such written agreement relates to an Award), in a manner and to the extent it deems necessary or expedient to make the Plan or Award fully effective.
(iii) To settle all controversies regarding the Plan and Awards granted under it.
9
(iv) To accelerate the time at which an Award may first be exercised or the time during which an Award or any part thereof will vest, notwithstanding the terms set forth in the Award Agreement or any written agreement between the Company or any Affiliate and a Participant that is approved by the Board.
(v) To prohibit the exercise of any Option, Stock Appreciation Right or other exercisable Award pending stock dividend, stock split, combination or exchange of Shares, merger, consolidation or other distribution (other than normal cash dividends) of Company assets to stockholders, or any other change affecting the Shares or the Share price, including any Corporate Transaction, for reasons of administrative convenience for the period of time the Board determines is necessary or advisable in connection with the transaction.
(vi) To suspend or terminate the Plan at any time. Any suspension, or a termination, of the Plan will not Materially Impair rights and obligations of any Award granted while the Plan is in effect except with the consent of the affected Participant.
(vii) To amend the Plan in any respect the Board deems necessary or advisable; provided, however, that stockholder approval will be required for any amendment to the extent required by Applicable Law. Except as provided above, rights of a Participant under any Award granted before amendment of the Plan will not be Materially Impaired except with the consent of the affected Participant.
(viii) To submit any amendment to the Plan for stockholder approval.
(ix) To approve forms of Award Agreements for use under the Plan and to amend the terms of any one or more Awards, including, but not limited to, amendments to provide terms more favorable to the Participant than previously provided in the Award Agreement, subject to any specified limits in the Plan that are not subject to Board discretion; provided however, that, a Participant’s rights under any Award will not be Materially Impaired except with the consent of the affected Participant.
(x) Generally, to exercise such powers and to perform such acts as the Board deems necessary or expedient to promote the best interests of the Company and that are not in conflict with the provisions of the Plan or outstanding Awards.
(xi) To adopt procedures and sub-plans as are necessary or appropriate to permit and facilitate participation in the Plan by, or take advantage of specific tax treatment for Awards granted to, Employees, Directors or Consultants who are non-U.S. nationals or employed outside the United States (provided that Board approval will not be necessary for immaterial modifications to the Plan or any Award Agreement to ensure or facilitate compliance with the laws, rules, or regulations of the relevant non-U.S. jurisdiction).
(xii) To effect, at any time and from time to time, subject to the consent of any Participant whose Award is Materially Impaired by such action, (1) the reduction of the exercise price (or strike price) of any outstanding Option or Stock Appreciation Right; (2) the cancellation of any outstanding Option or Stock Appreciation Right and the grant in substitution therefor of (A) a new Option, Stock Appreciation Right, Restricted Stock Award, Restricted Stock Unit Award or Other Award, under the Plan or any similar award under another equity plan of the Company, covering the same or a different number of Shares, (B) cash and/or (C) other valuable consideration (as determined by the Board); or (3) any other action that is treated as a repricing under generally accepted accounting principles.
10
(c) Delegation to Committee.
(i) General. The Board may delegate some or all of the administration of the Plan to a Committee or Committees. If administration of the Plan is delegated to a Committee, the Committee will have, in connection with the administration of the Plan, the powers theretofore possessed by the Board that have been delegated to the Committee, including the power to delegate to another Committee or a subcommittee of the Committee any of the administrative powers the Committee is authorized to exercise (and references in this Plan to the Board will thereafter be to the Committee or subcommittee), subject, however, to such resolutions, not inconsistent with the provisions of the Plan, as may be adopted from time to time by the Board. Each Committee may retain the authority to concurrently administer the Plan with the Committee or subcommittee to which it has delegated its authority hereunder and may, at any time, revest in such Committee some or all of the powers previously delegated. The Board may retain the authority to concurrently administer the Plan with any Committee and may, at any time, revest in the Board some or all of the powers previously delegated.
(ii) Rule 16b-3 Compliance. To the extent an Award is intended to qualify for the exemption from Section 16(b) of the Exchange Act that is available under Rule 16b-3 of the Exchange Act, the Award will be granted by the Board or a Committee that consists solely of two or more Non-Employee Directors, as determined under Rule 16b-3(b)(3) of the Exchange Act and thereafter any action establishing or modifying the terms of the Award will be approved by the Board or a Committee meeting such requirements to the extent necessary for such exemption to remain available.
(d) Effect of Board’s Decision. All determinations, interpretations and constructions made by the Board or any Committee in good faith will not be subject to review by any person and will be final, binding, and conclusive on all persons.
(e) Delegation to Other Person or Body. The Board or any Committee may delegate to one or more persons or bodies the authority to administer the Plan as permitted by Applicable Law, including, without limitation, Sections 152 and 157 of the Delaware General Corporation Law.
(f) Delegation of Authority for Day-to-Day Administration. Except to the extent prohibited by Applicable Law, the Plan Administrator is delegated the day-to-day administration of the Plan and the functions assigned to Plan Administrator by the Board or Committees. Any delegation to the Plan Administrator may be revoked at any time.
8. TAX WITHHOLDING
(a) Withholding Authorization. As a condition to acceptance of any Award under the Plan, a Participant authorizes withholding from payroll and any other amounts payable to such Participant, and otherwise agrees to make adequate arrangements to satisfy the Tax-Related Items withholding obligations, if any, of the Company and/or an Affiliate that arise in connection with the grant, vesting, exercise or settlement of such Award, as applicable. Accordingly, a Participant may not be able to exercise an Award even though the Award is vested, and the Company shall have no obligation to issue Shares subject to an Award, unless and until such obligations are satisfied.
(b) Satisfaction of Withholding Obligation. To the extent permitted by the terms of an Award Agreement, the Company may, in its sole discretion, satisfy any Tax-Related Items withholding obligation relating to an Award by any of the following means or by a combination of such means: (i) causing the Participant to tender a cash payment; (ii) withholding Shares otherwise issuable to the Participant in connection with the Award; (iii) withholding cash from an Award settled in cash; (iv) withholding payment from any amounts otherwise payable to the Participant; (v) by allowing a Participant to effectuate a “cashless exercise” pursuant to a program adopted by the Company in connection with the Plan; or (vi) by such other method as may be set forth in the Award Agreement and otherwise permitted by Applicable Law.
11
(c) No Obligation to Notify or Minimize Losses or Taxes; No Liability to Claims. Except as required by Applicable Law, the Company has no duty or obligation to any Participant to advise the holder of an Award as to the time or manner of exercising the Award. Furthermore, the Company has no duty or obligation to warn or otherwise advise the holder of an Award of a pending termination or expiration of the Award or a possible period in which the Award may not be exercised. The Company has no duty or obligation to minimize the tax consequences of an Award to the holder of such Award and will not be liable to any holder of an Award for any adverse tax consequences to such holder in connection with an Award. As a condition to accepting an Award under the Plan, each Participant (i) agrees to not make any claim against the Company, or any of its Officers, Directors, Employees or Affiliates related to tax liabilities arising from such Award or other Company compensation and (ii) acknowledges that such Participant was advised to consult with the Participant’s own personal tax, financial and other legal advisors regarding the tax consequences of the Award and has either done so or knowingly and voluntarily declined to do so. Additionally, each Participant acknowledges any Option or Stock Appreciation Right granted under the Plan is exempt from Section 409A only if the exercise or strike price is at least equal to the “fair market value” of the underlying Shares on the date of grant as determined by the U.S. Internal Revenue Service and there is no other impermissible deferral of compensation associated with the Award. Additionally, as a condition to accepting an Option or Stock Appreciation Right granted under the Plan, each Participant agrees to not make any claim against the Company, or any of its Officers, Directors, Employees or Affiliates in the event that the U.S. Internal Revenue Service asserts that the exercise price or strike price is less than the “fair market value” of the underlying Shares on the date of grant as subsequently determined by the U.S. Internal Revenue Service.
(d) Withholding Indemnification. The Company and its Affiliate may withhold or account for Tax-Related Items by considering statutory or other withholding rates, including minimum or maximum rates applicable in a Participant’s jurisdiction. In the event of over-withholding, the Participant may receive a refund of any over-withheld amount in cash (with no entitlement to the equivalent in Shares) or, if not refunded, the Participant may seek a refund from the local tax authorities. In the event of under-withholding, the Participant may be required to pay any additional Tax-Related Items directly to the applicable tax authority or to the Company and/or its Affiliate. As a condition to accepting an Award under the Plan, in the event that the amount of the Company’s and/or its Affiliate’s withholding obligation in connection with such Award was greater than the amount actually withheld by the Company and/or its Affiliates, each Participant agrees to indemnify and hold the Company and/or its Affiliates harmless from any failure by the Company and/or its Affiliates to withhold the proper amount. Further, if the obligation for Tax-Related Items is satisfied by withholding in Shares, for tax purposes, the Participant will be deemed to have been issued the full number of Shares subject to the Award, notwithstanding that a number of the Shares is held back solely for the purpose of paying the Tax-Related Items.
9. MISCELLANEOUS.
(a) Source of Shares. The Shares issuable under the Plan will be Shares of authorized but unissued or reacquired Common Stock, including Shares repurchased by the Company on the open market or otherwise.
12
(b) Corporate Action Constituting Grant of Awards. Corporate action constituting a grant by the Company of an Award to any Participant will be deemed completed as of the date of such corporate action, unless otherwise determined by the Board, regardless of when the instrument, certificate, or letter evidencing the Award is communicated to, or actually received or accepted by, the Participant. In the event that the corporate records (e.g., Board consents, resolutions or minutes) documenting the corporate action approving the grant contain terms (e.g., exercise price, vesting schedule or number of Shares) that are inconsistent with those in the Award Agreement or related grant documents as a result of a clerical error in the Award Agreement or related grant documents, the corporate records will control and the Participant will have no legally binding right to the incorrect term in the Award Agreement or related grant documents.
(c) Stockholder Rights. No Participant will be deemed to be the holder of, or to have any of the rights of a holder with respect to, any Shares subject to such Award unless and until (i) the Participant has satisfied all requirements for exercise of the Award pursuant to its terms, if applicable, and (ii) the issuance of the Shares subject to the Award is reflected in the records of the Company.
(d) No Employment or Other Service Rights. Nothing in the Plan, any Award Agreement or any other instrument executed thereunder or in connection with any Award granted pursuant thereto will confer upon any Participant any right to continue to serve the Company or an Affiliate in the capacity in effect at the time the Award was granted or affect the right of the Company or an Affiliate to terminate at will (unless otherwise required under Applicable Law) and without regard to any future vesting opportunity that a Participant may have with respect to any Award (i) the employment of an Employee with or without notice and with or without Cause, (ii) the service of a Consultant pursuant to the terms of such Consultant’s agreement with the Company or an Affiliate, or (iii) the service of a Director pursuant to the Bylaws of the Company or an Affiliate, and any applicable provisions of the corporate law of the U.S. state or non-U.S. jurisdiction in which the Company or the Affiliate is incorporated, as the case may be. Further, nothing in the Plan, any Award Agreement or any other instrument executed thereunder or in connection with any Award will constitute any promise or commitment by the Company or an Affiliate regarding the fact or nature of future positions, future work assignments, future compensation or any other term or condition of employment or service or confer any right or benefit under the Award or the Plan unless such right or benefit has specifically accrued under the terms of the Award Agreement and/or Plan.
(e) Change in Time Commitment. In the event a Participant’s regular level of time commitment in the performance of the Participant’s services for the Company and any Affiliates is reduced (for example, and without limitation, if the Participant is an Employee of the Company and the Employee has a change in status from a full-time Employee to a part-time Employee or takes an extended leave of absence) after the date of grant of any Award to the Participant, the Board may determine, to the extent permitted by Applicable Law, to (i) make a corresponding reduction in the number of Shares or cash amount subject to any portion of such Award that is scheduled to vest or become payable after the date of such change in time commitment, and (ii) in lieu of or in combination with a reduction, extend the vesting or payment schedule applicable to the Award. In the event of any extension or reduction, the Participant will not be deemed to be Materially Impaired, no consent from the Participant will be required, and the Participant will have no right with respect to any portion of the Award that is so reduced.
(f) Execution of Additional Documents. As a condition to accepting an Award under the Plan, the Participant agrees to execute any additional documents or instruments necessary or desirable, as determined in the Plan Administrator’s sole discretion, to carry out the purposes or intent of the Award, or facilitate compliance with securities and/or other regulatory requirements, in each case at the Plan Administrator’s request.
(g) Electronic Delivery and Participation. Any reference herein or in an Award Agreement to a “written” agreement or document will include any agreement or document delivered electronically, filed publicly at www.sec.gov (or any successor website thereto) or posted on the Company’s intranet (or other shared electronic medium controlled by the Company to which the Participant has access). By accepting any Award the Participant consents to receive documents by electronic delivery and to participate in the Plan through any on-line electronic system established and maintained by the Plan Administrator or another third party selected by the Plan Administrator. The form of delivery of any Shares (e.g., a stock certificate or electronic entry evidencing such Shares) shall be determined by the Company.
13
(h) Clawback/Recovery. Each Award granted under the Plan will be subject to reduction, cancellation, forfeiture, or recoupment in accordance with any clawback policy of the Company that is in effect as of the date the Award is granted and any clawback policy that the Company is required to adopt pursuant to the listing standards of any national securities exchange or association on which the Company’s securities are listed or as is otherwise required by the Dodd-Frank Wall Street Reform and Consumer Protection Act or other Applicable Law. In addition, the Board may impose such other clawback, recovery or recoupment provisions in an Award Agreement as the Board determines necessary or appropriate, including but not limited to a reacquisition right in respect of previously acquired Shares or cash upon the occurrence of Cause. No recovery of compensation under a clawback policy will be an event giving rise to a Participant’s right to voluntarily terminate employment upon a “resignation for good reason,” or for a “constructive termination” or any similar term under any plan of or agreement with the Company.
(i) Securities Law Compliance. A Participant will not be issued any Shares in respect of an Award unless either (i) the Shares are registered under the Securities Act or (ii) the Company has determined that such issuance would be exempt from the registration requirements of the Securities Act. Each Award also must comply with other Applicable Law governing the Award, and a Participant will not receive Shares if the Company determines that the receipt of Shares would not be in compliance with Applicable Law. If required by the Company, issuance will be further subject to the approval of counsel for the Company with respect to such compliance. If the Company determines it to be impossible or impractical to obtain authority from any regulatory body having jurisdiction or to complete or comply with the requirements of any Applicable Laws, registration or other qualification of the Shares under any state, federal or foreign law or under the rules and regulations of the U.S. Securities and Exchange Commission, the stock or share exchange on which Shares of the same class are then listed, or any other governmental or regulatory body, which authority, registration, qualification or rule compliance is deemed by the Company’s counsel to be necessary or advisable for the issuance and sale of any Shares hereunder, the Company will be relieved of any liability regarding the failure to issue or sell such Shares as to which such authority, registration, qualification or rule compliance was not obtained and the Board reserves the authority, without the consent of a Participant, to terminate or cancel Awards with or without consideration in such a situation.
As a condition to the exercise of an Award, the Company may require the person exercising such Award to represent and warrant at the time of any exercise that the Shares are being purchased only for investment and without any present intention to sell or distribute such Shares if, in the opinion of counsel for the Company, such a representation is required.
The inability of the Company to obtain authority from any regulatory body having jurisdiction or to complete or comply with the requirements of any registration or other qualification of the Shares under any U.S. state or federal law or non-U.S. law or under the rules and regulations of the Securities and Exchange Commission, the stock exchange on which Shares of the same class are then listed, or any other governmental or regulatory body, which authority, registration, qualification or rule compliance is deemed by the Company’s counsel to be necessary or advisable for the issuance and sale of any Shares hereunder, will relieve the Company of any liability in respect of the failure to issue or sell such Shares as to which such requisite authority, registration, qualification or rule compliance will not have been obtained.
14
(j) Transfer or Assignment of Awards; Issued Shares. Except as expressly provided in the Plan or the applicable Award Agreement, Awards granted under the Plan may not be transferred or assigned by the Participant. After the vested Shares subject to an Award have been issued, or in the case of a Restricted Stock Award and similar Awards, after the issued Shares have vested, the holder of the Shares is free to assign, hypothecate, donate, encumber or otherwise dispose of any interest in those Shares provided that any such actions are in compliance with the provisions herein, the terms of the Trading Policy and Applicable Law.
(k) Effect on Other Employee Benefit Plans. The value of any Award granted under the Plan, as determined upon grant, vesting or settlement, shall not be included as compensation, earnings, salaries, or other similar terms used when calculating any Participant’s benefits under any employee benefit plan sponsored by the Company or any Affiliate, except as such plan otherwise expressly provides. The Company expressly reserves its rights to amend, modify, or terminate any of the Company’s or any Affiliate’s employee benefit plans.
(l) Deferrals. To the extent permitted by Applicable Law, the Board, in its sole discretion, may determine that the delivery of Shares or the payment of cash, upon the exercise, vesting or settlement of all or a portion of an Award may be deferred and may also establish programs and procedures for deferral elections to be made by Participants.
(m) Section 409A. Unless otherwise expressly provided for in an Award Agreement, the Plan and Award Agreements will be interpreted to the greatest extent possible in a manner that makes the Plan and the Awards granted hereunder exempt from Section 409A, and, to the extent not so exempt, in compliance with the requirements of Section 409A. If the Board determines that any Award granted hereunder is not exempt from and is therefore subject to Section 409A, the Award Agreement evidencing such Award will incorporate the terms and conditions necessary to avoid the consequences specified in Section 409A(a)(1) of the Code, and to the extent an Award Agreement is silent on terms necessary for compliance, such terms are hereby incorporated by reference into the Award Agreement. Notwithstanding anything to the contrary in this Plan (and unless the Award Agreement specifically provides otherwise), if the Shares are publicly traded, and if a Participant holding an Award that constitutes “deferred compensation” under Section 409A is a “specified employee” for purposes of Section 409A, no distribution or payment of any amount that is due because of a “separation from service” (as defined in Section 409A without regard to alternative definitions thereunder) will be issued or paid before the date that is six months and one day following the date of the Participant’s “separation from service” or, if earlier, the date of the Participant’s death, unless the distribution or payment can be made in a manner that complies with Section 409A, and any amounts so deferred will be paid in a lump sum on the day after the six month period elapses, with the balance paid thereafter on the original schedule.
(n) Choice of Law. This Plan and any controversy arising out of or relating to this Plan shall be governed by, and construed in accordance with, the internal laws of the State of Delaware, without regard to conflict of law principles that would result in any application of any law other than the law of the State of Delaware.
10. SEVERABILITY.
If all or any part of the Plan, any Award Agreement, or any written agreement between a Participant and the Company or any Affiliate of the Company that is approved by the Board (to the extent such written agreement relates to an Award) is declared by any court or governmental authority to be unlawful or invalid, such unlawfulness or invalidity shall not invalidate any portion of the Plan or such Award Agreement or written agreement not declared to be unlawful or invalid. Any Section of the Plan or any Award Agreement or such written agreement (or part of such a Section) so declared to be unlawful or invalid shall, if possible, be construed in a manner which will give effect to the terms of such Section or part of a Section to the fullest extent possible while remaining lawful and valid.
15
11. TERMINATION OF THE PLAN.
The Board may suspend or terminate the Plan at any time. No Incentive Stock Options may be granted after the tenth anniversary of the Effective Date. No Awards may be granted under the Plan while the Plan is suspended or after it is terminated.
12. DEFINITIONS.
As used in the Plan, the following definitions apply to the capitalized terms indicated below:
(a) “Acquiring Entity” means the surviving or acquiring corporation (or its parent company) in connection with a Corporate Transaction.
(b) “Affiliate” means, at the time of determination, any “parent” or “subsidiary” of the Company as such terms are defined in Rule 405 promulgated under the Securities Act. The Board may determine the time or times at which “parent” or “subsidiary” status is determined within the foregoing definition.
(c) “Applicable Law” means the Code and any applicable U.S. and non-U.S. securities, exchange, control, tax, federal, state, material local or municipal or other law, statute, constitution, principle of common law, resolution, ordinance, code, edict, decree, rule, listing rule, regulation, judicial decision, ruling or requirement issued, enacted, adopted, promulgated, implemented or otherwise put into effect by or under the authority of any Governmental Body (including under the authority of any applicable self-regulating organization such as the Nasdaq Stock Market, New York Stock Exchange, or the Financial Industry Regulatory Authority).
(d) “Award” means any right to receive Shares or cash granted under the Plan (including an Incentive Stock Option, a Nonstatutory Stock Option, a Restricted Stock Award, a Restricted Stock Unit Award, a Stock Appreciation Right, a Performance Award or any Other Award).
(e) “Award Agreement” means a written or electronic agreement between the Company and a Participant evidencing the terms and conditions of an Award. The Award Agreement generally consists of the Grant Notice and the agreement containing the written summary of the general terms and conditions applicable to the Award and which is provided, including through electronic means, to a Participant along with the Grant Notice.
(f) “Board” means the Board of Directors of the Company (or its designee). Any decision or determination made by the Board shall be a decision or determination that is made in the sole discretion of the Board (or its designee), and such decision or determination shall be final and binding on all Participants.
(g) “Capital Stock” means each and every class of common stock of the Company, regardless of the number of votes per share.
(h) “Capitalization Adjustment” means any change that is made in, or other events that occur with respect to, the Common Stock subject to the Plan or subject to any Award after the Effective Date without the receipt of consideration by the Company through merger, consolidation, reorganization, recapitalization, reincorporation, stock dividend, dividend in property other than cash, large nonrecurring cash dividend, stock split, reverse stock split, liquidating dividend, split-up, spin-off, combination of Shares or other securities of the Company, reclassification, repurchase or exchange of Shares or other securities of the Company, change in corporate structure or any similar equity restructuring transaction, as that term is used in Statement of Financial Accounting Standards Board Accounting Standards Codification Topic 718 (or any successor thereto). Notwithstanding the foregoing, the conversion of any convertible securities of the Company will not be treated as a Capitalization Adjustment.
16
(i) “Cause” has the meaning ascribed to such term in any written agreement between a Participant and the Company or any Affiliate of the Company that is approved by the Board defining such term and, in the absence of such agreement, such term means, with respect to a Participant, the occurrence of any of the following events: (i) the Participant’s dishonest statements or acts with respect to the Company or any Affiliate of the Company, or any current or prospective customers, suppliers, vendors or other third parties with which such entity does business; (ii) the Participant’s commission of (A) a felony or (B) any misdemeanor involving moral turpitude, deceit, dishonesty or fraud, or in each case the equivalent in any relevant jurisdiction; (iii) the Participant’s failure to perform the Participant’s assigned duties and responsibilities to the reasonable satisfaction of the Company or any Affiliate of the Company which failure continues, in the reasonable judgment of the Company, after written notice given to the Participant by the Company or any Affiliate of the Company; (iv) the Participant’s gross negligence, willful misconduct or insubordination with respect to the Company or any Affiliate of the Company; or (v) the Participant’s material violation of any provision of any agreement(s) between the Participant and the Company or any Affiliate of the Company relating to noncompetition, nonsolicitation, nondisclosure and/or assignment of inventions. The determination that a termination of the Participant’s Continuous Service is either for Cause or without Cause will be made by the Board with respect to Participants who are executive officers of the Company and by the Company’s Chief Executive Officer with respect to Participants who are not executive officers of the Company. Any determination by the Company that the Continuous Service of a Participant was terminated with or without Cause for the purposes of outstanding Awards held by such Participant will have no effect upon any determination of the rights or obligations of the Company or any Affiliate of the Company or such Participant for any other purpose.
(j) “Change in Control” or “Change of Control” means the occurrence, in a single transaction or in a series of related transactions, of any one or more of the following events:
(i) any Exchange Act Person becomes the Owner, directly or indirectly, of securities of the Company representing more than 50% of the combined voting power of the Company’s then outstanding securities other than by virtue of a merger, consolidation, or similar transaction. Notwithstanding the foregoing, a Change in Control shall not be deemed to occur (A) on account of the acquisition of securities of the Company directly from the Company, (B) on account of the acquisition of securities of the Company by an investor, any affiliate thereof or any other Exchange Act Person that acquires the Company’s securities in a transaction or series of related transactions the primary purpose of which is to obtain financing for the Company through the issuance of equity securities, or (C) solely because the level of Ownership held by any Exchange Act Person (the “Subject Person”) exceeds the designated percentage threshold of the outstanding voting securities as a result of a repurchase or other acquisition of voting securities by the Company reducing the number of Shares outstanding, provided that if a Change in Control would occur (but for the operation of this sentence) as a result of the acquisition of voting securities by the Company, and after such share acquisition, the Subject Person becomes the Owner of any additional voting securities that, assuming the repurchase or other acquisition had not occurred, increases the percentage of the then outstanding voting securities Owned by the Subject Person over the designated percentage threshold, then a Change in Control shall be deemed to occur;
(ii) there is consummated a merger, consolidation or similar transaction involving (directly or indirectly) the Company and, immediately after the consummation of such merger, consolidation or similar transaction, the stockholders of the Company immediately prior thereto do not Own, directly or indirectly, either (A) outstanding voting securities representing more than 50% of the combined outstanding voting power of the Acquiring Entity in such merger, consolidation or similar transaction or (B) more than 50% of the combined outstanding voting power of the parent of the Acquiring Entity in such merger, consolidation or similar transaction, in each case in substantially the same proportions as their Ownership of the outstanding voting securities of the Company immediately prior to such transaction;
17
(iii) there is consummated a sale, lease, exclusive license or other disposition of all or substantially all of the consolidated assets of the Company and its Subsidiaries, other than a sale, lease, license or other disposition of all or substantially all of the consolidated assets of the Company and its Subsidiaries to an Entity, more than 50% of the combined voting power of the voting securities of which are Owned by stockholders of the Company in substantially the same proportions as their Ownership of the outstanding voting securities of the Company immediately prior to such sale, lease, license or other disposition; or
(iv) individuals who, on the Effective Date, are members of the Board (the “Incumbent Board”) cease for any reason to constitute at least a majority of the members of the Board; provided, however, that if the appointment or election (or nomination for election) of any new Board member was approved or recommended by a majority vote of the members of the Incumbent Board then still in office, such new member shall, for purposes of this Plan, be considered as a member of the Incumbent Board.
Notwithstanding the foregoing or any other provision of this Plan, (A) the term Change in Control shall not include a sale of assets, merger or other transaction effected exclusively for the purpose of changing the domicile of the Company, (B) the definition of Change in Control (or any analogous term) in an individual written agreement between the Company or any Affiliate and the Participant that is approved by the Board shall supersede the foregoing definition with respect to Awards subject to such agreement; provided, however, that if no definition of Change in Control or any analogous term is set forth in such an individual written agreement, the foregoing definition shall apply, and (C) with respect to any nonqualified deferred compensation that becomes payable on account of the Change in Control, the transaction or event described in clause (i), (ii), (iii), or (iv) must constitute a Section 409A Change in Control if required in order for the payment not to violate Section 409A of the Code.
(k) “Code” means the U.S. Internal Revenue Code of 1986, as amended, including any applicable regulations and guidance thereunder.
(l) “Committee” means the Compensation Committee and any other committee of one or more Directors to whom authority has been delegated by the Board or Compensation Committee in accordance with the Plan.
(m) “Common Stock” means the common stock of the Company.
(n) “Company” means Electra Therapeutics, Inc., a Delaware corporation, and any successor corporation thereto.
(o) “Compensation Committee” means the Compensation Committee of the Board.
(p) “Consultant” means any person, including an advisor, who is (i) engaged by the Company or an Affiliate to render consulting or advisory services and is compensated for such services, or (ii) serving as a member of the board of directors of an Affiliate and is compensated for such services. However, service solely as a Director, or payment of a fee for such service, will not cause a Director to be considered a “Consultant” for purposes of the Plan. Notwithstanding the foregoing, a person is treated as a Consultant under this Plan only if a Form S-8 Registration Statement under the Securities Act is available to register either the offer or the sale of the Company’s securities to such person.
18
(q) “Continuous Service” means that the Participant’s service with the Company or an Affiliate, whether as an Employee, Director, or Consultant, is not interrupted or terminated. A change in the capacity in which the Participant renders service to the Company or an Affiliate as an Employee, Director or Consultant or a change in the Entity for which the Participant renders such service, provided that there is no interruption or termination of the Participant’s service with the Company or an Affiliate, will not terminate a Participant’s Continuous Service; provided, however, that if the Entity for which a Participant is rendering services ceases to qualify as an Affiliate, as determined by the Board, such Participant’s Continuous Service will be considered to have terminated on the date such Entity ceases to qualify as an Affiliate. For example, a change in status from an Employee of the Company to a Consultant of an Affiliate or to a Director will not constitute an interruption of Continuous Service. To the extent permitted by Applicable Law, the Board or the chief executive officer of the Company, in that party’s sole discretion, may determine whether Continuous Service will be considered interrupted in the case of (i) any leave of absence approved by the Company or an Affiliate, including sick leave, military leave or any other personal leave, or (ii) transfers between the Company, an Affiliate, or their successors. Notwithstanding the foregoing, a leave of absence will be treated as Continuous Service for purposes of vesting in an Award only to such extent as may be provided in the Company’s leave of absence policy, in the written terms of any leave of absence agreement or policy applicable to the Participant, or as otherwise required by Applicable Law. In addition, to the extent required for exemption from or compliance with Section 409A, the determination of whether there has been a termination of Continuous Service will be made, and such term will be construed, in a manner that is consistent with the definition of “separation from service” as defined in Section 409A (without regard to any alternative definition thereunder).
(r) “Corporate Transaction” means the consummation, in a single transaction or in a series of related transactions, of any one or more of the following events:
(i) a sale or other disposition of all or substantially all, as determined by the Board, of the consolidated assets of the Company and its Subsidiaries;
(ii) a sale or other disposition of at least 50% of the outstanding securities of the Company;
(iii) a merger, consolidation or similar transaction following which the Company is not the surviving corporation; or
(iv) a merger, consolidation or similar transaction following which the Company is the surviving corporation but the Shares outstanding immediately preceding the merger, consolidation or similar transaction are converted or exchanged by virtue of the merger, consolidation, or similar transaction into other property, whether in the form of securities, cash or otherwise.
Notwithstanding the foregoing or any other provision of this Plan, (A) the term Corporate Transaction shall not include a sale of assets, merger or other transaction effected exclusively for the purpose of changing the domicile of the Company, (B) the definition of Corporate Transaction (or any analogous term) in an individual written agreement between the Company or any Affiliate and the Participant that is approved by the Board shall supersede the foregoing definition with respect to Awards subject to such agreement; provided, however, that if no definition of Corporate Transaction or any analogous term is set forth in such an individual written agreement, the foregoing definition shall apply, and (C) with respect to any nonqualified deferred compensation that becomes payable on account of the Corporate Transaction, the transaction or event described in clause (i), (ii), (iii), or (iv) must constitute a Section 409A Change in Control if required in order for the payment not to violate Section 409A of the Code.
19
(s) “Director” means a member of the Board.
(t) “Disability” means, with respect to a Participant, such Participant is unable to engage in any substantial gainful activity by reason of any medically determinable physical or mental impairment which can be expected to result in death or which has lasted or can be expected to last for a continuous period of not less than 12 months, as provided in Section 22(e)(3) of the Code, and will be determined by the Board on the basis of such medical evidence as the Board deems warranted under the circumstances.
(u) “Effective Date” means the date that is immediately prior to the IPO Date.
(v) “Employee” means any person employed by the Company or an Affiliate. However, service solely as a Director, or payment of a fee for such services, will not cause a Director to be considered an “Employee” for purposes of the Plan.
(w) “Employer” means the Company or the Affiliate that employs the Participant.
(x) “Entity” means a corporation, partnership, limited liability company or other entity.
(y) “Exchange Act” means the U.S. Securities Exchange Act of 1934, as amended, and the rules and regulations promulgated thereunder.
(z) “Exchange Act Person” means any natural person, Entity or “group” (within the meaning of Section 13(d) or 14(d) of the Exchange Act), except that “Exchange Act Person” will not include (i) the Company or any Subsidiary of the Company, (ii) any employee benefit plan of the Company or any Subsidiary of the Company or any trustee or other fiduciary holding securities under an employee benefit plan of the Company or any Subsidiary of the Company, (iii) an underwriter temporarily holding securities pursuant to a registered public offering of such securities, (iv) an Entity Owned, directly or indirectly, by the stockholders of the Company in substantially the same proportions as their Ownership of stock of the Company, or (v) any natural person, Entity or “group” (within the meaning of Section 13(d) or 14(d) of the Exchange Act) that, as of the Effective Date, is the Owner, directly or indirectly, of securities of the Company representing more than 50% of the combined voting power of the Company’s then outstanding securities.
(aa) “Fair Market Value” means, as of any date, unless otherwise determined by the Board, the value of the Common Stock (as determined on a per Share or aggregate basis, as applicable) determined as follows:
(i) If the Common Stock is listed on any established stock exchange or traded on any established market, the Fair Market Value will be the closing sales price for such stock as quoted on such exchange or market (or the exchange or market with the greatest volume of trading in the Common Stock) on the date of determination, as reported in a source the Board deems reliable.
(ii) If there is no closing sales price for the Common Stock on the date of determination, then the Fair Market Value will be the closing selling price on the last preceding date for which such quotation exists.
(iii) For purposes of any Awards granted on or before the effective date of the Registration Statement (as defined below), the Fair Market Value will be the initial price to the public as set forth in the final prospectus included within the registration statement on Form S-1 filed with the Securities and Exchange Commission for the initial public offering of the Common Stock (the “Registration Statement”).
20
(iv) In the absence of such markets for the Common Stock, or if otherwise determined by the Board, the Fair Market Value will be determined by the Board in good faith and in a manner that complies with Sections 409A and 422 of the Code.
(bb) “Fully Diluted Capital Stock” means the number of shares of Capital Stock, determined as of the applicable date of measurement, equal to the sum of (i) the number of shares of Capital Stock issued and outstanding and (ii) the number of shares of Capital Stock issuable upon the exercise, conversion, or exchange of all then-outstanding pre-funded warrants, convertible Capital Stock or indebtedness, exchangeable Capital Stock or indebtedness, or other rights exercisable for or convertible into, directly or indirectly, shares of Capital Stock (whether vested or unvested), but excluding the number of shares of Capital Stock issuable upon the exercise of then-outstanding options and the shares reserved and available for issuance under the Company’s then-existing equity incentive plan(s) and employee stock purchase plan(s), if any.
(cc) “Governmental Body” means any: (i) nation, state, commonwealth, canton, province, territory, county, municipality, district or other jurisdiction of any nature; (ii) U.S. or non-U.S. federal, state, local, municipal, or other government; (iii) governmental or regulatory body, or quasi-governmental body of any nature (including any governmental division, department, administrative agency or bureau, commission, authority, instrumentality, official, ministry, fund, foundation, center, organization, unit, body or Entity and any court or other tribunal, and for the avoidance of doubt, any tax authority) or other body exercising similar powers or authority; or (iv) self-regulatory organization (including the Nasdaq Stock Market, New York Stock Exchange, and the Financial Industry Regulatory Authority).
(dd) “Grant Notice” means the notice provided to a Participant that the Participant has been granted an Award under the Plan and that includes the name of the Participant, the type of Award, the date of grant of the Award, number of Shares subject to the Award or potential cash payment right, (if any), the vesting schedule for the Award (if any) and other key terms applicable to the Award.
(ee) “Incentive Stock Option” means an option granted pursuant to Section 4 of the Plan that is intended to be, and qualifies as, an “incentive stock option” within the meaning of Section 422 of the Code.
(ff) “IPO Date” means the date of the underwriting agreement between the Company and the underwriter(s) managing the initial public offering of the Common Stock, pursuant to which the Common Stock is priced for the initial public offering.
(gg) “Materially Impair” means any amendment to the terms of the Award that materially adversely affects the Participant’s rights under the Award. A Participant’s rights under an Award will not be deemed to have been Materially Impaired by any such amendment if the Board, in its sole discretion, determines that the amendment, taken as a whole, does not materially impair the Participant’s rights. For example, the following types of amendments to the terms of an Award do not Materially Impair the Participant’s rights under the Award: (i) imposition of reasonable restrictions on the minimum number of Shares subject to an Option or Stock Appreciation Right that may be exercised; (ii) to maintain the qualified status of the Award as an Incentive Stock Option under Section 422 of the Code; (iii) to change the terms of an Incentive Stock Option in a manner that disqualifies, impairs or otherwise affects the qualified status of the Award as an Incentive Stock Option under Section 422 of the Code; (iv) to clarify the manner of exemption from, or to bring the Award into compliance with or qualify it for an exemption from, Section 409A; or (v) to comply with other Applicable Laws.
21
(hh) “Non-Employee Director” means a Director who either (i) is not a current employee or officer of the Company or an Affiliate, does not receive compensation, either directly or indirectly, from the Company or an Affiliate for services rendered as a consultant or in any capacity other than as a Director (except for an amount as to which disclosure would not be required under Item 404(a) of Regulation S-K promulgated pursuant to the Securities Act (“Regulation S-K”)), does not possess an interest in any other transaction for which disclosure would be required under Item 404(a) of Regulation S-K, and is not engaged in a business relationship for which disclosure would be required pursuant to Item 404(b) of Regulation S-K; or (ii) is otherwise considered a “non-employee director” for purposes of Rule 16b-3.
(ii) “Nonstatutory Stock Option” means any option granted pursuant to Section 4 of the Plan that does not qualify as an Incentive Stock Option.
(jj) “Officer” means a person who is an officer of the Company within the meaning of Section 16 of the Exchange Act.
(kk) “Option” means an Incentive Stock Option or a Nonstatutory Stock Option to purchase Shares granted pursuant to the Plan.
(ll) “Other Award” means an award valued in whole or in part by reference to, or otherwise based on, Shares, including the appreciation in value thereof (e.g., options or stock rights with an exercise price or strike price less than 100% of the Fair Market Value at the time of grant) that is not an Incentive Stock Option, Nonstatutory Stock Option, Stock Appreciation Right, Restricted Stock Award, Restricted Stock Unit Award or Performance Award.
(mm) “Own,” “Owned,” “Owner,” or “Ownership” means that a person or Entity will be deemed to “Own,” to have “Owned,” to be the “Owner” of, or to have acquired “Ownership” of securities if such person or Entity, directly or indirectly, through any contract, arrangement, understanding, relationship or otherwise, has or shares voting power, which includes the power to vote or to direct the voting, with respect to such securities.
(nn) “Participant” means an Employee, Director, or Consultant to whom an Award is granted pursuant to the Plan or, if applicable, such other person who holds an outstanding Award.
(oo) “Performance Award” means an Award that may vest or may be exercised or a cash award that may vest or become earned and paid contingent upon the attainment during a Performance Period of certain Performance Goals and which is granted under the terms and conditions of Section 5(b) pursuant to such terms as are approved by the Board. In addition, to the extent permitted by Applicable Law and set forth in the applicable Award Agreement, the Board may determine that cash may be used in payment of Performance Awards. Performance Awards that are settled in cash are not required to be valued in whole or in part by reference to, or otherwise based on, Shares.
(pp) “Performance Criteria” means one or more criteria that the Board will select for purposes of establishing the Performance Goals for a Performance Period. The Performance Criteria that will be used to establish such Performance Goals may be based on any one of, or combination of, the following as determined by the Board: earnings (including earnings per Share and net earnings); earnings before interest, taxes and depreciation; earnings before interest, taxes, depreciation and amortization; total stockholder return; relative stockholder return; return on equity or average stockholder’s equity; return on assets, investment, or capital employed; stock price; margin (including gross margin); income (before or after taxes); operating income; operating income after taxes; pre-tax profit; operating cash flow; sales, annual recurring revenue or revenue targets; increases in revenue or product revenue; expenses and cost reduction
22
goals; improvement in or attainment of working capital levels; economic value added (or an equivalent metric); market share; cash flow; cash flow per Share; Share price performance; debt reduction; customer satisfaction; stockholders’ equity; capital expenditures; debt levels; operating profit or net operating profit; workforce diversity; growth of net income or operating income; billings; financing; regulatory milestones; stockholder liquidity; corporate governance and compliance; intellectual property; personnel matters; progress of internal research; progress of partnered programs; partner satisfaction; budget management; partner or collaborator achievements; internal controls, including those related to the U.S. Sarbanes-Oxley Act of 2002; investor relations, analysts and communication; implementation or completion of projects or processes; employee retention; number of users, including unique users; strategic partnerships or transactions (including in-licensing and out-licensing of intellectual property); establishing relationships with respect to the marketing, distribution and sale of the Company’s products; supply chain achievements; co-development, co-marketing, profit sharing, joint venture or other similar arrangements; individual performance goals; corporate development and planning goals; and other measures of performance selected by the Board or Committee whether or not listed herein.
(qq) “Performance Goals” means, for a Performance Period, one or more goals established by the Board for the Performance Period based upon the Performance Criteria. Performance Goals may be based on a Company-wide basis, with respect to one or more business units, divisions, Affiliates, or business segments, and in either absolute terms or relative to the performance of one or more comparable companies or the performance of one or more relevant indices. Unless specified otherwise by the Board (i) in the Award Agreement at the time the Award is granted or (ii) in such other document setting forth the Performance Goals at the time the Performance Goals are established, the Board will appropriately make adjustments in the method of calculating the attainment of Performance Goals for a Performance Period as follows: (1) to exclude restructuring and/or other nonrecurring charges; (2) to exclude exchange rate effects; (3) to exclude the effects of changes to generally accepted accounting principles; (4) to exclude the effects of any statutory adjustments to corporate tax rates; (5) to exclude the effects of items that are “unusual” in nature or occur “infrequently” as determined under generally accepted accounting principles; (6) to exclude the dilutive effects of acquisitions or joint ventures; (7) to assume that any business divested by the Company achieved performance objectives at targeted levels during the balance of a Performance Period following such divestiture; (8) to exclude the effect of any change in the outstanding Shares by reason of any stock dividend or split, stock repurchase, reorganization, recapitalization, merger, consolidation, spin-off, combination or exchange of Shares or other similar corporate change, or any distributions to common stockholders other than regular cash dividends; (9) to exclude the effects of stock based compensation and the award of bonuses under the Company’s bonus plans; (10) to exclude costs incurred in connection with potential acquisitions or divestitures that are required to be expensed under generally accepted accounting principles; and (11) to exclude the goodwill and intangible asset impairment charges that are required to be recorded under generally accepted accounting principles. In addition, the Board may establish or provide for other adjustment items in the Award Agreement at the time the Award is granted or in such other document setting forth the Performance Goals at the time the Performance Goals are established. In addition, the Board retains the discretion to reduce or eliminate the compensation or economic benefit due upon attainment of Performance Goals and to define the manner of calculating the Performance Criteria it selects to use for such Performance Period. Partial achievement of the specified criteria may result in the payment or vesting corresponding to the degree of achievement as specified in the Award Agreement or the written terms of a Performance Award.
(rr) “Performance Period” means the period of time selected by the Board over which the attainment of one or more Performance Goals will be measured for the purpose of determining a Participant’s right to vesting or exercise of an Award. Performance Periods may be of varying and overlapping duration, at the sole discretion of the Board.
23
(ss) “Plan” means this Electra Therapeutics, Inc. 2026 Equity Incentive Plan, as amended from time to time.
(tt) “Plan Administrator” means the person, persons, or third-party administrator designated by the Company to administer the day-to-day operations of the Plan and the Company’s other equity incentive programs.
(uu) “Post-Termination Exercise Period” means the period following termination of a Participant’s Continuous Service within which an Option or Stock Appreciation Right is exercisable, as specified in Section 4(h).
(vv) “Prior Plan” means the Electra Therapeutics, Inc. 2022 Stock Plan, as amended from time to time.
(ww) “Restricted Stock Award” means an Award of Shares which is granted pursuant to the terms and conditions of Section 5(a).
(xx) “Returning Shares” means Shares subject to outstanding awards granted under the Prior Plan and that following the Effective Date: (A) are not issued because such award or any portion thereof expires or otherwise terminates without all of the Shares covered by such award having been issued; (B) are not issued because such award or any portion thereof is settled in cash; (C) are forfeited back to or repurchased by the Company because of the failure to meet a contingency or condition required for the vesting of such award; (D) are withheld or reacquired to satisfy the exercise, strike or purchase price related to the award; or (E) are withheld or reacquired to satisfy a tax withholding obligation related to the award.
(yy) “Restricted Stock Unit Award” means an Award of restricted stock units representing the right to receive an issuance of Shares which is granted pursuant to the terms and conditions of Section 5(a).
(zz) “Rule 16b-3” means Rule 16b-3 promulgated under the Exchange Act or any successor to Rule 16b-3, as in effect from time to time.
(aaa) “Rule 405” means Rule 405 promulgated under the Securities Act.
(bbb) “Section 409A” means Section 409A of the Code and the regulations and other guidance thereunder.
(ccc) “Section 409A Change in Control” means a change in the ownership or effective control of the Company, or in the ownership of a substantial portion of the Company’s assets, as provided in Section 409A(a)(2)(A)(v) of the Code and U.S. Treasury Regulations Section 1.409A-3(i)(5) (without regard to any alternative definition thereunder).
(ddd) “Securities Act” means the U.S. Securities Act of 1933, as amended.
(eee) “Share” means a share of the Common Stock, as adjusted for a Capitalization Adjustment.
(fff) “Share Reserve” means the number of Shares available for issuance under the Plan as set forth in Section 2(a).
(ggg) “Stock Appreciation Right” means a right to receive the appreciation on Shares that is granted pursuant to the terms and conditions of Section 4.
24
(hhh) “Subsidiary” means, with respect to the Company, (i) any corporation of which more than 50% of the outstanding Capital Stock having ordinary voting power to elect a majority of the board of directors of such corporation (irrespective of whether, at the time, stock of any other class or classes of such corporation will have or might have voting power by reason of the happening of any contingency) is at the time, directly or indirectly, Owned by the Company, and (ii) any partnership, limited liability company or other entity in which the Company has a direct or indirect interest (whether in the form of voting or participation in profits or capital contribution) of more than 50%.
(iii) “Tax-Related Items” means any income tax, social insurance, payroll tax, fringe benefit tax, payment on account or other tax-related items arising out of or in relation to a Participant’s participation in the Plan and legally applicable or deemed applicable to the Participant.
(jjj) “Ten Percent Stockholder” means a person who Owns (or is deemed to Own pursuant to Section 424(d) of the Code) stock possessing more than 10% of the total combined voting power of all classes of stock of the Company or any “parent corporation” or “subsidiary corporation” thereof (as such terms are defined in Sections 424(e) and (f) of the Code).
(kkk) “Trading Policy” means the Company’s policy permitting certain individuals to sell Company Shares only during certain “window” periods or otherwise restricts the ability of certain individuals to transfer or encumber Shares, as in effect from time to time.
25
Exhibit 10.4
ELECTRA THERAPEUTICS, INC.
2026 EMPLOYEE STOCK PURCHASE PLAN
ADOPTED BY THE BOARD OF DIRECTORS: AUGUST 19, 2026
APPROVED BY THE STOCKHOLDERS: SEPTEMBER 10, 2026
| 1. | GENERAL; PURPOSE. |
(a) The Plan provides a means by which Eligible Employees of the Company and certain Designated Companies may be given an opportunity to purchase Shares. The Plan permits the Company to grant a series of Purchase Rights to Eligible Employees under an Employee Stock Purchase Plan. In addition, the Plan permits the Company to grant a series of Purchase Rights to Eligible Employees who do not meet the requirements of an Employee Stock Purchase Plan.
(b) The Plan includes two components: a 423 Component and a Non-423 Component. The Company intends (but makes no undertaking or representation to maintain) the 423 Component to qualify as an Employee Stock Purchase Plan. The provisions of the 423 Component, accordingly, will be construed in a manner that is consistent with the requirements of Section 423 of the Code. In addition, this Plan authorizes grants of Purchase Rights under the Non-423 Component that do not meet the requirements of an Employee Stock Purchase Plan. Except as otherwise provided in the Plan or determined by the Board, the Non-423 Component will operate and be administered in the same manner as the 423 Component. In addition, the Company may make separate Offerings which vary in terms (provided that such terms are not inconsistent with the provisions of the Plan or the requirements of an Employee Stock Purchase Plan to the extent the Offering is made under the 423 Component), and the Company will designate which Designated Company is participating in each separate Offering.
(c) The Company, by means of the Plan, seeks to retain the services of Eligible Employees, to secure and retain the services of new Employees and to provide incentives for such persons to exert maximum efforts for the success of the Company and its Related Corporations.
| 2. | ADMINISTRATION. |
(a) The Board will administer the Plan unless and until the Board delegates administration of the Plan to a Committee or Committees, as provided in Section 2(c).
(b) The Board will have the power, subject to, and within the limitations of, the express provisions of the Plan:
(i) To determine eligibility, how and when Purchase Rights will be granted, and the provisions of each Offering (which need not be identical). Without limiting the generality of, and consistent with, the foregoing, the Board is authorized to determine that, to the extent permitted by U.S. Treasury Regulation Section 1.423-2(f), the terms of a Purchase Right granted under the Plan or an Offering to citizens or residents of a non-U.S. jurisdiction will be less favorable than the terms of Purchase Rights granted under the Plan or the same Offering to employees resident solely in the United States.
(ii) To designate from time to time (A) which Related Corporations will be eligible to participate in the Plan as Designated 423 Companies, (B) which Related Corporations or Affiliates will be eligible to participate in the Plan as Designated Non-423 Companies, (C) which Affiliates or Related Corporations may be excluded from participation in the Plan, and (D) which Designated Companies will participate in each separate Offering (to the extent that the Company makes separate Offerings).
(iii) To construe and interpret the Plan and Purchase Rights, and to establish, amend and revoke rules and regulations for its administration. The Board, in the exercise of this power, may correct any defect, omission or inconsistency in the Plan, in a manner and to the extent it deems necessary or expedient to make the Plan fully effective.
(iv) To settle all controversies regarding the Plan and Purchase Rights granted under the Plan.
(v) To suspend or terminate the Plan at any time as provided in Section 11.
(vi) To amend the Plan at any time as provided in Section 11.
(vii) Generally, to exercise such powers and to perform such acts as it deems necessary or expedient to promote the best interests of the Company, its Related Corporations and Affiliates, and to carry out the intent that the Plan be treated as an Employee Stock Purchase Plan with respect to the 423 Component.
(viii) To adopt such rules, procedures, sub-plans, and appendices as are necessary or appropriate for the administration of the Plan (including, without limitation, such rules, procedures, sub-plans, and appendices as are necessary or appropriate to permit or facilitate participation in the Plan by Employees who are non-U.S. nationals or employed or located outside the United States, the terms of which rules, procedures, sub-plans and appendices may take precedence over other provisions of this Plan, with the exception of Section 3(a) hereof, but unless otherwise superseded by the terms of such rules, procedures, sub-plans, or appendices, the provisions of this Plan will govern the operation of such rules, procedures, sub-plans, and appendices). Without limiting the generality of, and consistent with, the foregoing, the Board specifically is authorized to adopt rules, procedures, and sub-plans regarding, without limitation, eligibility to participate in the Plan, the definition of eligible “earnings,” handling and making of Contributions, establishment of bank or trust accounts to hold Contributions, payment of interest (which will apply to all Participants in the relevant Offerings under the 423 Component, except to the extent otherwise permitted by U.S. Treasury Regulation Section 1.423-2(f)), conversion of local currency, obligations to pay payroll tax, determination of beneficiary designation requirements, withholding procedures and handling of share issuances, any of which may vary according to applicable requirements, and which, if applicable to a Designated Non-423 Company, do not have to comply with the requirements of Section 423 of the Code. Unless otherwise determined by the Board, the Eligible Employees eligible to participate in each sub-plan will participate in a separate Offering under the 423 Component, or if the terms would not qualify under the 423 Component, in the Non-423 Component, in either case unless such designation would cause the 423 Component to violate the requirements of Section 423 of the Code.
(c) The Board may delegate some or all of the administration of the Plan to a Committee or Committees. If administration is delegated to a Committee, the Committee will have, in connection with the administration of the Plan, the powers theretofore possessed by the Board that have been delegated to the Committee, including the power to delegate to a subcommittee any of the administrative powers the Committee is authorized to exercise (and references in this Plan and any applicable Offering Document to the Board will thereafter be to the Committee or subcommittee), subject, however, to such resolutions, not inconsistent with the provisions of the Plan, as may be adopted from time to time by the Board. Further, to the extent not prohibited by Applicable Law, the Board or Committee may, from time to time, delegate some or all of its authority under the Plan to one or more officers of the Company or other persons or groups of persons as it deems necessary, appropriate or advisable under conditions or limitations that it may set at or after the time of the delegation. The Board may retain the authority to concurrently administer the Plan with the Committee (or its delegate) and may, at any time, revest in the Board some or all of the powers previously delegated. Whether or not the Board has delegated administration of the Plan to a Committee (or a delegate of the Committee), the Board will have the final power to determine all questions of policy and expediency that may arise in the administration of the Plan.
2
(d) All determinations, interpretations and constructions made by the Board will not be subject to review by any person and will be final, binding and conclusive on all persons.
| 3. | SHARES SUBJECT TO THE PLAN. |
(a) Subject to the provisions of Section 10(a) relating to Capitalization Adjustments, the maximum number of Shares that may be issued under the Plan will not exceed 611,682 Shares, plus the number of Shares that are automatically added on January 1st of each year for a period of up to ten years, commencing on January 1, 2027 and ending on (and including) January 1, 2036, in an amount equal to the lesser of (x) 1% of the Fully Diluted Capital Stock outstanding on December 31st of the preceding calendar year, or (y) 1,835,046 Shares. Notwithstanding the foregoing, the Board may act prior to the first day of any calendar year to provide that there will be no January 1st increase in the share reserve for such calendar year or that the increase in the share reserve for such calendar year will be a lesser number of Shares than would otherwise occur pursuant to the preceding sentence. For the avoidance of doubt, up to the maximum number of Shares reserved under this Section 3(a) may be used to satisfy purchases of Shares under the 423 Component and any remaining portion of such maximum number of Shares may be used to satisfy purchases of Shares under the Non-423 Component.
(b) If any Purchase Right granted under the Plan terminates without having been exercised in full, the Shares not purchased under such Purchase Right will again become available for issuance under the Plan.
(c) The stock purchasable under the Plan will be Shares of authorized but unissued or reacquired Common Stock, including Shares repurchased by the Company on the open market or otherwise.
| 4. | GRANT OF PURCHASE RIGHTS; OFFERING. |
(a) The Board may from time to time grant or provide for the grant of Purchase Rights to Eligible Employees under an Offering (consisting of one or more Purchase Periods) on an Offering Date or Offering Dates selected by the Board. Each Offering will be in such form and will contain such terms and conditions as the Board will deem appropriate, and with respect to the 423 Component, will comply with the requirement of Section 423(b)(5) of the Code that all Employees granted Purchase Rights will have the same rights and privileges. The terms and conditions of an Offering will be incorporated by reference into the Plan and treated as part of the Plan. The provisions of separate Offerings need not be identical, but each Offering will include (through incorporation of the provisions of this Plan by reference in the document comprising the Offering or otherwise) the period during which the Offering will be effective, which period will not exceed 27 months beginning with the Offering Date, and the substance of the provisions contained in Sections 5 through 8, inclusive.
(b) If a Participant has more than one Purchase Right outstanding under the Plan, unless such Participant otherwise indicates in forms delivered to the Company or a third party designated by the Company (each, a “Company Designee”): (i) each form will apply to all of such Participant’s Purchase Rights under the Plan, and (ii) a Purchase Right with a lower exercise price (or an earlier-granted Purchase Right, if different Purchase Rights have identical exercise prices) will be exercised to the fullest possible extent before a Purchase Right with a higher exercise price (or a later-granted Purchase Right if different Purchase Rights have identical exercise prices) will be exercised.
3
(c) The Board will have the discretion to structure an Offering so that if the Fair Market Value of a Share on the first Trading Day of a new Purchase Period within that Offering is less than or equal to the Fair Market Value of a Share on the Offering Date for that Offering, then (i) that Offering will terminate immediately as of that first Trading Day, and (ii) the Participants in such terminated Offering will be automatically enrolled in a new Offering beginning on the first Trading Day of such new Purchase Period.
| 5. | ELIGIBILITY. |
(a) Purchase Rights may be granted only to Employees of the Company or, as the Board may designate in accordance with Section 2(b), to Employees of a Related Corporation or an Affiliate. Except as provided in Section 5(b) or as required by Applicable Law, an Employee will not be eligible to be granted Purchase Rights unless, on the Offering Date, the Employee has been in the employ of the Company, the Related Corporation or the Affiliate, as the case may be, for such continuous period preceding such Offering Date as the Board may (unless prohibited by Applicable Law) require, but in no event will the required period of continuous employment be equal to or greater than two years. In addition, the Board may provide (unless prohibited by Applicable Law) that no Employee will be eligible to be granted Purchase Rights under the Plan unless, on the Offering Date, such Employee’s customary employment with the Company, the Related Corporation or the Affiliate is more than 20 hours per week and more than five months per calendar year or such other criteria as the Board may determine consistent with Section 423 of the Code with respect to the 423 Component. The Board may also exclude (unless prohibited by Applicable Law) from participation in the Plan or any Offering (i) any “highly compensated employees” (within the meaning of Section 423(b)(4)(D) of the Code) of the Company, a Related Corporation or an Affiliate or (ii) any such highly compensated employees with compensation above a certain level or who are officers or subject to the disclosure requirements of Section 16(a) of the Exchange Act, provided that the exclusion in this clause (ii) is applied with respect to each Offering under the 423 Component in an identical manner to all highly compensated individuals of the entity whose employees are participating in that Offering. Each exclusion will be applied with respect to an Offering under the 423 Component in a manner complying with U.S. Treasury Regulation Section 1.423-2(e)(2)(ii). Such exclusions may be applied with respect to an Offering under the Non-423 Component without regard to the limitations of U.S. Treasury Regulation Section 1.423-2.
(b) The Board may provide that each person who, during the course of an Offering, first becomes an Eligible Employee will, on a date or dates specified in the Offering which coincides with the day on which such person becomes an Eligible Employee or which occurs thereafter, receive a Purchase Right under that Offering, which Purchase Right will thereafter be deemed to be a part of that Offering. Such Purchase Right will have the same characteristics as any Purchase Rights originally granted under that Offering, as described herein, except that:
(i) the date on which such Purchase Right is granted will be the “Offering Date” of such Purchase Right for all purposes, including determination of the exercise price of such Purchase Right;
(ii) the period of the Offering with respect to such Purchase Right will begin on its Offering Date and end coincident with the end of such Offering; and
(iii) the Board may provide that if such person first becomes an Eligible Employee within a specified period of time before the end of the Offering, the individual will not receive any Purchase Right under that Offering.
4
(c) No Employee will be eligible for the grant of any Purchase Rights under the 423 Component if, immediately after any such Purchase Rights are granted, such Employee owns stock possessing five percent or more of the total combined voting power or value of all classes of stock of the Company or of any Related Corporation. For purposes of this Section 5(c), the rules of Section 424(d) of the Code will apply in determining the stock ownership of any Employee, and stock which such Employee may purchase under all outstanding Purchase Rights and options will be treated as stock owned by such Employee.
(d) As specified by Section 423(b)(8) of the Code, an Eligible Employee may be granted Purchase Rights under the 423 Component only if such Purchase Rights, together with any other rights granted under all Employee Stock Purchase Plans of the Company and any Related Corporations, do not permit such Eligible Employee’s rights to purchase stock of the Company or any Related Corporation to accrue at a rate which, when aggregated, exceeds U.S. $25,000 of Fair Market Value of such stock (determined at the time such rights are granted, and which, with respect to the Plan, will be determined as of their respective Offering Dates) for each calendar year in which such rights are outstanding at any time.
(e) Officers of the Company and any Designated Company, if they are otherwise Eligible Employees, will be eligible to participate in Offerings under the Plan. Notwithstanding the foregoing, the Board may (unless prohibited by Applicable Law) provide in an Offering that Employees who are highly compensated Employees within the meaning of Section 423(b)(4)(D) of the Code will not be eligible to participate.
(f) Notwithstanding anything in this Section 5 to the contrary, in the case of an Offering under the Non-423 Component, an Eligible Employee (or group of Eligible Employees) may be excluded from participation in the Plan or an Offering if the Board has determined, in its sole discretion, that participation of such Eligible Employee(s) is not advisable or practical for any reason.
| 6. | PURCHASE RIGHTS; PURCHASE PRICE. |
(a) On each Offering Date, each Eligible Employee, pursuant to an Offering made under the Plan, will be granted a Purchase Right to purchase up to that number of Shares purchasable either with a percentage of earnings (as such concept is defined in the Offering Document) or with a maximum dollar amount, but in either case as so specified by the Board in the Offering Document, during the period that begins on the Offering Date (or such later date as the Board determines for a particular Offering) and ends on the date stated in the Offering, which date will be no later than the end of the Offering.
(b) The Board will establish one or more Purchase Dates during an Offering on which Purchase Rights granted for that Offering will be exercised and Shares will be purchased in accordance with such Offering.
(c) In connection with each Offering made under the Plan, the Board may specify (i) a maximum number of Shares that may be purchased by any Participant on any Purchase Date during such Offering, (ii) a maximum aggregate number of Shares that may be purchased by all Participants pursuant to such Offering and/or (iii) a maximum aggregate number of Shares that may be purchased by all Participants on any Purchase Date under the Offering. If the aggregate purchase of Shares issuable upon exercise of Purchase Rights granted under the Offering would exceed any such maximum aggregate number, then, in the absence of any Board action otherwise, a pro rata (based on each Participant’s accumulated Contributions) allocation of the Shares (rounded down to the nearest whole Share) available will be made in as nearly a uniform manner as will be practicable and equitable.
(d) The purchase price of Shares acquired pursuant to Purchase Rights will be specified by the Board prior to commencement of an Offering and will not be less than the lesser of:
5
(i) an amount equal to 85% of the Fair Market Value of the Shares on the Offering Date; or
(ii) an amount equal to 85% of the Fair Market Value of the Shares on the applicable Purchase Date.
| 7. | PARTICIPATION; WITHDRAWAL; TERMINATION. |
(a) An Eligible Employee may elect to participate in an Offering and authorize payroll deductions as the means of making Contributions by completing and delivering to the Company or a Company Designee, within the time specified in the Offering, an enrollment form approved by the Board. The enrollment form will specify the amount of Contributions not to exceed the maximum amount specified by the Board. Each Participant’s Contributions will be credited to a bookkeeping account for such Participant under the Plan and will be deposited with the general funds of the Company except where Applicable Law requires that Contributions be held separately or deposited with a third party. If permitted in the Offering, a Participant may begin such Contributions with the first practicable payroll occurring on or after the Offering Date (or, in the case of a payroll date that occurs after the end of the prior Offering but before the Offering Date of the next new Offering, Contributions from such payroll will be included in the new Offering). If permitted in the Offering, a Participant may thereafter reduce (including to zero) or increase such Participant’s Contributions. If payroll deductions are impermissible or problematic under Applicable Law or if specifically provided in the Offering and to the extent permitted by Section 423 of the Code with respect to the 423 Component, in addition to or instead of making Contributions by payroll deductions, a Participant may make Contributions through payment by cash, check or wire transfer prior to a Purchase Date.
(b) During an Offering, a Participant may cease making Contributions and withdraw from the Offering by delivering to the Company or a Company Designee a withdrawal form approved by the Board. The Company may impose a deadline before a Purchase Date for withdrawing. Upon such withdrawal, such Participant’s Purchase Right in that Offering will immediately terminate, and the Company will distribute as soon as practicable to such Participant all of such Participant’s accumulated but unused Contributions. A Participant’s withdrawal from that Offering will have no effect upon such Participant’s eligibility to participate in any other Offerings under the Plan, but such Participant will be required to deliver a new enrollment form (in accordance with Section 7(a)) to participate in subsequent Offerings.
(c) A Participant’s Purchase Rights granted pursuant to any Offering under the Plan will terminate immediately if the Participant either (i) is no longer an Employee for any reason or for no reason or (ii) is otherwise no longer eligible to participate in such Offering. The Company will distribute as soon as practicable to such individual all of such individual’s accumulated but unused Contributions.
(d) Unless otherwise determined by the Board, a Participant whose employment transfers or whose employment terminates with an immediate rehire (with no break in service) by or between the Company and a Designated Company or between Designated Companies will not be treated as having terminated employment for purposes of participating in the Plan or an Offering; however, if a Participant transfers from an Offering under the 423 Component to an Offering under the Non-423 Component, the exercise of the Participant’s Purchase Right will be qualified under the 423 Component only to the extent such exercise complies with Section 423 of the Code. If a Participant transfers from an Offering under the Non-423 Component to an Offering under the 423 Component, the exercise of the Purchase Right will remain non-qualified under the Non-423 Component for the remainder of the Offering. The Board may establish different and additional rules governing transfers between separate Offerings within the 423 Component and between Offerings under the 423 Component and Offerings under the Non-423 Component.
6
(e) During a Participant’s lifetime, Purchase Rights will be exercisable only by such Participant. Neither Contributions credited to a Participant’s account nor any Purchase Rights may be assigned, transferred, pledged, hypothecated, or otherwise disposed of in any way, except by will, by the laws of descent and distribution, or, if permitted by the Company and valid under Applicable Law, by a beneficiary designation as described in Section 9. Any such attempt at assignment, transfer, pledge, hypothecation, or other disposition will be without effect, except that the Company may treat such act as an election to withdraw funds from an Offering in accordance with Section 7(b).
(f) Unless otherwise specified in the Offering or required by Applicable Law, the Company will have no obligation to pay interest on Contributions.
| 8. | EXERCISE OF PURCHASE RIGHTS. |
(a) On each Purchase Date, each Participant’s accumulated Contributions will be applied to the purchase of Shares, up to the maximum number of Shares permitted by the Plan and the applicable Offering, at the purchase price specified in the Offering. No fractional Shares will be issued unless specifically provided for in the Offering.
(b) Unless otherwise provided in the Offering, if any amount of accumulated Contributions remains in a Participant’s account after the purchase of Shares and such remaining amount is less than the amount required to purchase one Share on the final Purchase Date of an Offering, then such remaining amount will be held in such Participant’s account for the purchase of Shares under the next Offering under the Plan, unless such Participant withdraws from or is not eligible to participate in such next Offering, in which case such amount will be distributed to such Participant after the final Purchase Date without interest (unless the payment of interest is otherwise required by Applicable Law). If the amount of Contributions remaining in a Participant’s account after the purchase of Shares is at least equal to the amount required to purchase one (1) whole Share on the final Purchase Date of an Offering, then such remaining amount will be distributed in full to such Participant after the final Purchase Date of such Offering without interest (unless otherwise required by Applicable Law).
(c) No Purchase Rights may be exercised to any extent unless the issuance of Shares pursuant to that exercise is in compliance with all applicable U.S. and non-U.S. federal, state and other securities, exchange control and other laws applicable to the Plan (including, but not limited to, the Shares being covered by an effective registration statement pursuant to the Securities Act). If on a Purchase Date the issuance of Shares would not be in such compliance, no Purchase Rights will be exercised on such Purchase Date, and, subject to Section 423 of the Code with respect to the 423 Component, the Purchase Date will be delayed until the issuance of Shares would be in compliance, except that the Purchase Date will in no event be more than 27 months from the Offering Date. If, on the Purchase Date, as delayed to the maximum extent permissible, the issuance of Shares is not in compliance with all Applicable Laws, as determined by the Company in its sole discretion, the Purchase Rights will not be exercised and all accumulated but unused Contributions for the applicable Offering will be distributed to the Participants without interest (unless the payment of interest is otherwise required by Applicable Law).
(d) As soon as reasonably practicable after each Purchase Date on which a purchase of Shares occurs, the Company will arrange the delivery to each Participant of the Shares purchased upon exercise of the Participant’s Purchase Right in a form determined by the Board (in its sole discretion) and pursuant to rules established by the Board. Shares to be delivered to a Participant under the Plan will be registered in the name of the Participant or, if so required under Applicable Laws, in the name of the Participant and the Participant’s spouse. The Company may permit or require that the Shares be deposited directly with a broker designated by the Company or to a designated agent of the Company, and the Company may utilize electronic or automated methods of share transfer. The Company may require that Shares be retained with the broker or agent for a designated period of time and/or may establish other procedures to permit tracking of disqualifying dispositions of the Shares.
7
| 9. | DESIGNATION OF BENEFICIARY. |
(a) The Board may, but is not obligated to, permit a Participant to submit a form designating a beneficiary who will receive any Shares and/or Contributions from the Participant’s account under the Plan if the Participant dies before such Shares and/or Contributions are delivered to the Participant. The Board may, but is not obligated to, permit the Participant to change such designation of beneficiary. Any such designation or change must be on a form approved by the Board. If a Participant is married and the designated beneficiary is not the spouse, spousal consent will be required for the designation or change to be effective.
(b) If a Participant dies, and in the absence of a valid beneficiary designation, the Company will deliver any Shares and/or Contributions to the executor or administrator of the estate of the Participant. If no executor or administrator has been appointed (to the knowledge of the Company), the Company, in its sole discretion, may deliver such Shares and/or Contributions, without interest (unless the payment of interest is otherwise required by Applicable Law), to the Participant’s spouse, dependents or relatives, or if no spouse, dependent or relative is known to the Company, then to such other person as the Company may designate.
| 10. | ADJUSTMENTS UPON CHANGES IN COMMON STOCK; CORPORATE TRANSACTIONS. |
(a) In the event of a Capitalization Adjustment, the Board will appropriately and proportionately adjust: (i) the class(es) and maximum number of securities subject to the Plan pursuant to Section 3(a), (ii) the class(es) and maximum number of securities by which the share reserve is to increase automatically each year pursuant to Section 3(a), (iii) the class(es) and number of securities subject to, and the purchase price applicable to outstanding Offerings and Purchase Rights, and (iv) the class(es) and number of securities that are the subject of the purchase limits under each ongoing Offering. The Board will make these adjustments, and its determination will be final, binding and conclusive.
(b) In the event of the proposed dissolution or liquidation of the Company, any Offering then in progress will be shortened by setting a Purchase Date (which will be before the date of the Company’s proposed dissolution or liquidation) and will terminate immediately prior to the consummation of the proposed dissolution or liquidation, unless provided otherwise by the Board. Prior to the new Purchase Date, the Board will provide written or electronic notice of the new Purchase Date to each Participant in the Offering.
(c) In the event of a Corporate Transaction, then: (i) any surviving corporation or acquiring corporation (or the surviving or acquiring corporation’s parent company) may assume or continue outstanding Purchase Rights or may substitute similar rights (including a right to acquire the same consideration paid to the stockholders in the Corporate Transaction) for outstanding Purchase Rights, or (ii) if any surviving or acquiring corporation (or its parent company) does not assume or continue such Purchase Rights or does not substitute similar rights for such Purchase Rights, then the Participants’ accumulated Contributions will be used to purchase Shares (rounded down to the nearest whole Share) within ten business days (or such other period specified by the Board) prior to the Corporate Transaction under the outstanding Purchase Rights, and the Purchase Rights will terminate immediately after such purchase.
8
| 11. | AMENDMENT, TERMINATION OR SUSPENSION OF THE PLAN. |
(a) The Board may amend the Plan at any time in any respect the Board deems necessary or advisable. However, except as provided in Section 10(a) relating to Capitalization Adjustments, stockholder approval will be required for any amendment of the Plan for which stockholder approval is required by Applicable Law. Notwithstanding anything in the Plan or any Offering Document to the contrary, the Board will be entitled to: (i) change the Offerings or Purchase Periods; (ii) designate separate Offerings; (iii) limit the frequency and/or number of changes in the amount withheld during an Offering; (iv) establish the exchange rate applicable to amounts withheld in a currency other than U.S. dollars; (v) permit Contributions in excess of the amount designated by a Participant in order to adjust for mistakes in the Company’s processing of properly completed Contribution elections; (vi) establish reasonable waiting and adjustment periods and/or accounting and crediting procedures to ensure that amounts applied toward the purchase of Shares for each Participant properly correspond with amounts withheld from the Participant’s Contributions; (vii) amend any outstanding Purchase Rights or clarify any ambiguities regarding the terms of any Offering to enable the Purchase Rights to qualify under and/or comply with Section 423 of the Code with respect to the 423 Component; and (viii) establish other limitations or procedures as the Board determines in its sole discretion advisable that are consistent with the Plan.
(b) The Board may suspend or terminate the Plan at any time. No Purchase Rights may be granted under the Plan while the Plan is suspended or after it is terminated. If the Plan is terminated, the Board, in its discretion, may elect to terminate all outstanding Offerings either immediately or upon completion of the purchase of Shares on the next Purchase Date (which may be sooner than originally scheduled, if determined by the Board in its discretion), or may elect to permit Offerings to expire in accordance with their terms (and subject to any adjustment pursuant to Section 10). If the Offerings are terminated prior to expiration, all amounts then credited to Participants’ accounts that have not been used to purchase Shares will be returned to the Participants (without interest thereon, except as otherwise required under Applicable Laws) as soon as administratively practicable.
| 12. | TAX QUALIFICATION; TAX WITHHOLDING. |
(a) Although the Company may endeavor to (i) qualify a Purchase Right for special tax treatment under the laws of the United States or jurisdictions outside of the United States or (ii) avoid adverse tax treatment, the Company makes no representation to that effect and expressly disavows any covenant to maintain special or to avoid unfavorable tax treatment, notwithstanding anything to the contrary in this Plan. The Company will be unconstrained in its corporate activities without regard to the potential negative tax impact on Participants.
(b) Each Participant will make arrangements, satisfactory to the Company and any applicable Related Corporation or Affiliate, to enable the Company, the Related Corporation or the Affiliate to fulfill any withholding obligation for Tax-Related Items. Without limitation to the foregoing, in the Company’s sole discretion and subject to Applicable Law, such withholding obligation may be satisfied in whole or in part by (i) withholding from the Participant’s salary or any other compensation due to the Participant from the Company, a Related Corporation or an Affiliate; (ii) withholding from the proceeds of the sale of Shares acquired under the Plan, either through a voluntary sale or a mandatory sale arranged by the Company; or (iii) any other method that is deemed acceptable by the Board and, with respect to the 423 Component, permitted by U.S. Treasury Regulation Section 1.423-2(f). The Company will not be required to issue any Shares under the Plan until such obligations are satisfied.
9
(c) The 423 Component is intended to be exempt from the application of Section 409A of the Code (or to the extent not exempt, to comply with Section 409A of the Code), and any ambiguities herein will be interpreted according to that intent. In furtherance of the foregoing and notwithstanding any provision in the Plan to the contrary, if the Board determines that a Purchase Right granted under the Plan may be subject to Section 409A of the Code or that any provision in the Plan would cause a Purchase Right under the Plan to be subject to Section 409A, the Board may amend the terms of the Plan and/or of an outstanding Purchase Right granted under the Plan, or take such other action the Board determines is necessary or appropriate, in each case, without the Participant’s consent, to exempt any outstanding Purchase Right or future Purchase Right that may be granted under the Plan from or to allow any such Purchase Right to comply with Section 409A of the Code, but only to the extent any such amendments or action by the Board would not violate Section 409A of the Code. Notwithstanding the foregoing, the Company will have no liability to a Participant or any other party if the Purchase Right under the Plan that is intended to be exempt from or compliant with Section 409A of the Code is not so exempt or compliant or for any action taken by the Board with respect thereto.
| 13. | EFFECTIVE DATE AND TERM OF PLAN. |
The Plan will become effective immediately prior to and contingent upon the IPO Date. No Purchase Rights will be exercised unless and until the Plan has been approved by the stockholders of the Company, which approval must be within 12 months before or after the date the Plan is adopted (or if required under Section 11(a) above, materially amended) by the Board. The Plan will continue in effect until terminated under Section 11.
| 14. | MISCELLANEOUS PROVISIONS. |
(a) The Company may use all Contributions received or held by it under the Plan for any corporate purpose, and the Company will not be obligated to segregate such Contributions except under Offerings or for Participants in the Non-423 Component for which Applicable Laws require that Contributions to the Plan by Participants be segregated from the Company’s general corporate funds and/or deposited with an independent third party, provided that, if such segregation or deposit with an independent third party is required by Applicable Laws, it will apply to all Participants in the relevant Offering under the 423 Component, except to the extent otherwise permitted by U.S. Treasury Regulation Section 1.423-2(f). Until Shares are issued, Participants will have only the rights of an unsecured creditor with respect to such Shares.
(b) A Participant will not be deemed to be the holder of, or to have any of the rights of a holder with respect to, Shares subject to Purchase Rights unless and until the Participant’s Shares acquired upon exercise of Purchase Rights are recorded in the books of the Company (or its transfer agent).
(c) The Plan and Offering do not constitute an employment contract. Nothing in the Plan or in the Offering will in any way alter the at will nature of a Participant’s employment or amend a Participant’s employment or service contract, as applicable, or be deemed to create in any way whatsoever any obligation on the part of any Participant to continue in the employ or service of the Company, a Related Corporation or an Affiliate, or on the part of the Company, a Related Corporation or an Affiliate to continue the employment or service of a Participant.
(d) The provisions of the Plan will be governed by the laws of the State of Delaware without resort to that state’s conflict of laws rules.
(e) If any particular provision of the Plan is found to be invalid or otherwise unenforceable, such provision will not affect the other provisions of the Plan, but the Plan will be construed in all respects as if such invalid provision were omitted.
10
(f) If any provision of the Plan does not comply with Applicable Law, such provision will be construed in such a manner as to comply with Applicable Law.
(g) Individual accounts will be maintained for each Participant in the Plan. Statements of account will be given to Participants at least annually, which statements will set forth the amounts of Contributions, the purchase price, the number of Shares purchased and the remaining cash balance, if any.
| 15. | DEFINITIONS. |
As used in the Plan, the following definitions will apply to the capitalized terms indicated below:
(a) “423 Component” means the part of the Plan, which excludes the Non-423 Component, pursuant to which Purchase Rights that satisfy the requirements for an Employee Stock Purchase Plan may be granted to Eligible Employees.
(b) “Affiliate” means any entity, other than a Related Corporation, whether now or subsequently established, which is at the time of determination, a “parent” or “subsidiary” of the Company as such terms are defined in Rule 405 promulgated under the Securities Act. The Board may determine the time or times at which “parent” or “subsidiary” status is determined within the foregoing definition.
(c) “Applicable Law” means the Code and any applicable U.S. and non-U.S. securities, exchange control, tax, federal, state, material local or municipal or other law, statute, constitution, principle of common law, resolution, ordinance, code, edict, decree, rule, listing rule, regulation, judicial decision, ruling or requirement issued, enacted, adopted, promulgated, implemented or otherwise put into effect by or under the authority of any Governmental Body (or under the authority of the New York Stock Exchange, Nasdaq Stock Market or the Financial Industry Regulatory Authority).
(d) “Board” means the Board of Directors of the Company.
(e) “Capital Stock” means each and every class of common stock of the Company, regardless of the number of votes per share.
(f) “Capitalization Adjustment” means any change that is made in, or other events that occur with respect to, the Common Stock subject to the Plan or subject to any Purchase Right after the date the Plan is adopted by the Board without the receipt of consideration by the Company through merger, consolidation, reorganization, recapitalization, reincorporation, stock dividend, dividend in property other than cash, large nonrecurring cash dividend, stock split, liquidating dividend, combination of shares, exchange of shares, change in corporate structure or other similar equity restructuring transaction, as that term is used in Financial Accounting Standards Board Accounting Standards Codification Topic 718 (or any successor thereto). Notwithstanding the foregoing, the conversion of any convertible securities of the Company will not be treated as a Capitalization Adjustment.
(g) “Code” means the U.S. Internal Revenue Code of 1986, as amended, including any applicable regulations and guidance thereunder.
(h) “Committee” means a committee of one or more members of the Board to whom authority has been delegated by the Board in accordance with Section 2(c).
(i) “Common Stock” means the common stock of the Company.
(j) “Company” means Electra Therapeutics, Inc., a Delaware corporation.
11
(k) “Contributions” means the payroll deductions, contributions made by Participants in case payroll deductions are impermissible or problematic under Applicable Law and other additional payments specifically provided for in the Offering that a Participant contributes to fund the exercise of a Purchase Right. A Participant may make additional payments into the Participant’s account if specifically provided for in the Offering, and then only if the Participant has not already had the maximum permitted amount withheld during the Offering through payroll deductions or other contributions and, with respect to the 423 Component, to the extent permitted by Section 423.
(l) “Corporate Transaction” means the consummation, in a single transaction or in a series of related transactions, of any one or more of the following events:
(i) a sale or other disposition of all or substantially all, as determined by the Board in its sole discretion, of the consolidated assets of the Company and its subsidiaries;
(ii) a sale or other disposition of more than 50% of the outstanding securities of the Company;
(iii) a merger, consolidation or similar transaction following which the Company is not the surviving corporation; or
(iv) a merger, consolidation or similar transaction following which the Company is the surviving corporation but the Shares outstanding immediately preceding the merger, consolidation or similar transaction are converted or exchanged by virtue of the merger, consolidation or similar transaction into other property, whether in the form of securities, cash or otherwise.
(m) “Designated 423 Company” means any Related Corporation selected by the Board as participating in the 423 Component.
(n) “Designated Company” means any Designated Non-423 Company or Designated 423 Company, provided, however, that at any given time, a Related Corporation participating in the 423 Component will not be a Related Corporation participating in the Non-423 Component.
(o) “Designated Non-423 Company” means any Related Corporation or Affiliate selected by the Board as participating in the Non-423 Component.
(p) “Director” means a member of the Board.
(q) “Eligible Employee” means an Employee who meets the requirements set forth in the document(s) governing the Offering for eligibility to participate in the Offering, provided that such Employee also meets the requirements for eligibility to participate set forth in the Plan.
(r) “Employee” means any person, including an Officer or Director, who is “employed” for purposes of Section 423(b)(4) of the Code by the Company or a Related Corporation or solely with respect to the Non-423 Component, an Affiliate. However, service solely as a Director, or payment of a fee for such services, will not cause a Director to be considered an “Employee” for purposes of the Plan.
(s) “Employee Stock Purchase Plan” means a plan that grants Purchase Rights intended to be options issued under an “employee stock purchase plan,” as that term is defined in Section 423(b) of the Code.
12
(t) “Exchange Act” means the U.S. Securities Exchange Act of 1934, as amended and the rules and regulations promulgated thereunder.
(u) “Fair Market Value” means, as of any date, the value of the Common Stock determined as follows:
(i) If the Common Stock is listed on any established stock exchange or traded on any established market, the Fair Market Value of a Share will be, unless otherwise determined by the Board, the closing sales price for such stock as quoted on such exchange or market (or the exchange or market with the greatest volume of trading in the Common Stock) on the date of determination, as reported in such source as the Board deems reliable. Unless otherwise provided by the Board, if there is no closing sales price for the Common Stock on the date of determination, then the Fair Market Value will be the closing sales price on the last preceding date for which such quotation exists.
(ii) In the absence of such markets for the Common Stock, the Fair Market Value will be determined by the Board in good faith in compliance with Applicable Laws and regulations and, to the extent applicable as determined in the sole discretion of the Board, in a manner that complies with Section 409A of the Code.
(iii) Notwithstanding the foregoing, for any Offering that commences on the IPO Date, the Fair Market Value of the Shares on the Offering Date will be the price per Share at which Shares are first sold to the public in the Company’s initial public offering as specified in the final prospectus for that initial public offering.
(v) “Fully Diluted Capital Stock” means the number of shares of Capital Stock, determined as of the applicable date of measurement, equal to the sum of (i) the number of shares of Capital Stock issued and outstanding and (ii) the number of shares of Capital Stock issuable upon the exercise, conversion, or exchange of all then-outstanding pre-funded warrants, convertible Capital Stock or indebtedness, exchangeable Capital Stock or indebtedness, or other rights exercisable for or convertible into, directly or indirectly, shares of Capital Stock (whether vested or unvested), but excluding the number of shares of Capital Stock issuable upon the exercise of then-outstanding options and the shares reserved and available for issuance under the Company’s then-existing equity incentive plan(s) and employee stock purchase plan(s), if any.
(w) “Governmental Body” means any: (i) nation, state, commonwealth, canton, province, territory, county, municipality, district or other jurisdiction of any nature; (ii) U.S. or non-U.S. federal, state, local, municipal or other government; (iii) governmental or regulatory body, or quasi-governmental body of any nature (including any governmental division, department, administrative agency or bureau, commission, authority, instrumentality, official, ministry, fund, foundation, center, organization, unit, body or entity and any court or other tribunal, and for the avoidance of doubt, any tax authority) or other body exercising similar powers or authority; or (iv) self-regulatory organization (including the New York Stock Exchange, the Nasdaq Stock Market and the Financial Industry Regulatory Authority).
(x) “IPO Date” means the date of the underwriting agreement between the Company and the underwriter(s) managing the initial public offering of the Common Stock, pursuant to which the Common Stock is priced for the initial public offering.
(y) “Non-423 Component” means the part of the Plan, which excludes the 423 Component, pursuant to which Purchase Rights that are not intended to satisfy the requirements for an Employee Stock Purchase Plan may be granted to Eligible Employees.
13
(z) “Offering” means the grant to Eligible Employees of Purchase Rights, with the exercise of those Purchase Rights automatically occurring at the end of one or more Purchase Periods. The terms and conditions of an Offering will generally be set forth in the “Offering Document” approved by the Board for that Offering.
(aa) “Offering Date” means a date selected by the Board for an Offering to commence.
(bb) “Officer” means a person who is an officer of the Company or a Related Corporation within the meaning of Section 16 of the Exchange Act.
(cc) “Participant” means an Eligible Employee who holds an outstanding Purchase Right.
(dd) “Plan” means this Electra Therapeutics, Inc. 2026 Employee Stock Purchase Plan, as amended from time to time, including both the 423 Component and the Non-423 Component.
(ee) “Purchase Date” means one or more dates during an Offering selected by the Board on which Purchase Rights will be exercised and on which purchases of Shares will be carried out in accordance with such Offering.
(ff) “Purchase Period” means a period of time specified within an Offering, generally beginning on the Offering Date or on the first Trading Day following a Purchase Date, and ending on a Purchase Date. An Offering may consist of one or more Purchase Periods.
(gg) “Purchase Right” means an option to purchase Shares granted pursuant to the Plan.
(hh) “Related Corporation” means any “parent corporation” or “subsidiary corporation” of the Company whether now or subsequently established, as those terms are defined in Sections 424(e) and (f), respectively, of the Code.
(ii) “Securities Act” means the U.S. Securities Act of 1933, as amended.
(jj) “Share” means a share of the Common Stock, as adjusted for a Capitalization Adjustment.
(kk) “Tax-Related Items” means any income tax, social insurance, payroll tax, fringe benefit tax, payment on account or other tax-related items arising out of or in relation to a Participant’s participation in the Plan, including, but not limited to, the exercise of a Purchase Right and the receipt of Shares or the sale or other disposition of Shares acquired under the Plan.
(ll) “Trading Day” means any day on which the exchange(s) or market(s) on which Shares are listed, including but not limited to the New York Stock Exchange, Nasdaq Global Select Market, the Nasdaq Global Market, the Nasdaq Capital Market or any successors thereto, is open for trading.
14
Exhibit 10.7
ELECTRA THERAPEUTICS, INC.
NON-EMPLOYEE DIRECTOR COMPENSATION POLICY
ADOPTED: AUGUST 19, 2026
EFFECTIVE DATE:
Each member of the Board of Directors (the “Board”) who is not also serving as an employee of or consultant to Electra Therapeutics, Inc. (the “Company”) or any of its subsidiaries (each such member, an “Eligible Director”) will receive the compensation described in this Non-Employee Director Compensation Policy (this “Policy”) for his or her Board service upon and following the date of the underwriting agreement between the Company and the underwriters managing the initial public offering of the Company’s common stock (the “Common Stock”), pursuant to which the Common Stock is priced in such initial public offering (the “Effective Date”). An Eligible Director may decline all or any portion of his or her or their compensation by giving notice to the Company prior to the date cash may be paid or equity awards are to be granted, as the case may be. This Policy is effective as of the Effective Date and may be amended at any time in the sole discretion of the Board.
Annual Cash Compensation
Commencing on the Effective Date, each Eligible Director will receive the cash compensation set forth below for service on the Board. Cash compensation amounts will be paid in equal quarterly installments, payable in arrears on the last day of each fiscal quarter in which the service occurred. If an Eligible Director joins the Board or a committee of the Board at a time other than effective as of the first day of a fiscal quarter, each annual retainer set forth below will be pro-rated based on days served in the applicable fiscal quarter, with the pro-rated amount paid for the first fiscal quarter in which the Eligible Director provides the service and regular full quarterly payments thereafter. All annual cash fees are vested upon payment.
| 1. | Annual Board Service Retainer: |
| a. | All Eligible Directors: $40,000 |
| b. | Non-Executive Chair of the Board (in addition to Eligible Director Service Retainer): $30,000 |
| 2. | Annual Committee Chair Service Retainer: |
| a. | Chair of the Audit Committee: $20,000 |
| b. | Chair of the Compensation Committee: $15,000 |
| c. | Chair of the Nominating and Corporate Governance Committee: $10,000 |
| 3. | Annual Committee Member Service Retainer (not applicable to Committee Chairs): |
| a. | Member of the Audit Committee: $10,000 |
| b. | Member of the Compensation Committee: $7,500 |
| c. | Member of the Nominating and Corporate Governance Committee: $5,000 |
1.
Equity Compensation
The equity compensation set forth below will be granted under the Company’s 2026 Equity Incentive Plan (the “Plan”), subject to the approval of the Plan by the Company’s stockholders. All stock options granted under this Policy will be nonstatutory stock options, with an exercise price per share equal to 100% of the Fair Market Value (as defined in the Plan) of the underlying Common Stock on the date of grant, and a term of ten years from the date of grant (subject to earlier termination in connection with a termination of service as provided in the Plan, provided that upon a termination of service other than for death, Disability or Cause (as each such term is defined in the Plan), the post-termination exercise period will be three months from the date of termination).
1. Initial Grant: For each Eligible Director who is first elected or appointed to the Board following the Effective Date, on the date of such Eligible Director’s initial election or appointment to the Board (or, if such date is not a market trading day, the first market trading day thereafter), the Eligible Director will be automatically, and without further action by the Board or the Compensation Committee of the Board, granted a stock option to purchase 22,632 shares of Common Stock (the “Initial Grant”). The shares subject to the Initial Grant shall vest in a series of 36 equal monthly installments over the three-year period measured from the date of grant, such that the Initial Grant shall be vested in full on the three-year anniversary of the grant date thereof, subject to the Eligible Director’s Continuous Service (as defined in the Plan) through each such vesting date.
2. Annual Grant: On the date of each annual stockholder meeting of the Company held after the Effective Date, each Eligible Director who continues to serve as a non-employee member of the Board following such stockholder meeting will be automatically, and without further action by the Board or the Compensation Committee of the Board, granted a stock option to purchase 45,264 shares of Common Stock (the “Annual Grant”). The shares subject to the Annual Grant will vest in full on the earlier of the date that is 12 months following the grant date thereof or the date of the Company’s next annual stockholder meeting, subject to the Eligible Director’s Continuous Service (as defined in the Plan) through such vesting date.
Change in Control
With respect to Awards (as defined in the Plan) granted to an Eligible Director while that individual was an Eligible Director, in the event of a Change in Control (as defined in the Plan), the Eligible Director will fully vest and have all other terms and conditions deemed to have been met and, if applicable, will have the right to exercise the Eligible Director’s outstanding Awards, unless specifically provided otherwise under the applicable award agreement or other written agreement authorized by the Board between the Eligible Director and the Company or any of its Affiliates (as defined in the Plan), as applicable.
2.
Expenses
The Company will reimburse an Eligible Director for ordinary, necessary and reasonable out-of-pocket travel expenses to cover in-person attendance at and participation in Board and committee meetings, and for other reasonable expenses incurred in connection with the Eligible Director’s service on the Board, including attendance at Board-related site visits, director education and training programs, industry conferences and any other activities undertaken at the request of the Board or the Company; provided, that such Eligible Director timely submit to the Company appropriate documentation substantiating such expenses in accordance with the Company’s travel and expense policy, as in effect from time to time.
Non-Employee Director Compensation Limit
Notwithstanding the foregoing, the aggregate value of all compensation granted or paid, as applicable, to any individual for service as a Non-Employee Director (as defined in the Plan) shall in no event exceed the limits set forth in Section 3(c) of the Plan.
Section 409A
In no event will cash compensation or expense reimbursement payments under this Policy be paid after the later of (i) the 15th day of the 3rd month following the end of the Company’s taxable year in which the compensation is earned or expenses are incurred, as applicable, or (ii) the 15th day of the 3rd month following the end of the calendar year in which the compensation is earned or expenses are incurred, as applicable, in compliance with the “short-term deferral” exception under Section 409A of the Internal Revenue Code of 1986, as amended, and the final regulations and guidance thereunder, as may be amended from time to time (together, “Section 409A”). It is the intent of this Policy that all payments hereunder be exempt from or otherwise comply with the requirements of Section 409A so that none of the compensation to be provided hereunder will be subject to the additional tax imposed under Section 409A, and any ambiguities or ambiguous terms herein will be interpreted to be so exempt or comply. In no event will the Company have any liability or obligation to reimburse, indemnify, or hold harmless an Eligible Director (or any other person) for any taxes or costs that may be imposed on or incurred by an Eligible Director (or any other person) as a result of Section 409A.
3.
Exhibit 10.14
STOCK RESTRICTION AGREEMENT
This Stock Restriction Agreement (the “Agreement”) is made as of , by and between Electra Therapeutics, Inc., a Delaware corporation (the “Company”) and [ ] (“Holder”). Certain capitalized terms used below are defined in the terms and conditions set forth in Exhibit A attached to this Agreement, which are incorporated by reference.
| Total Unvested Shares held by Holder: |
[ ] of shares of Series A Preferred Stock of the Company, including any securities into which such shares may convert (the “Unvested Shares”). | |
| Par Value: |
$0.0001 |
Vesting Schedule: Beginning on 1[September 16, 2026 (the “Vesting Commencement Date”), 1/13th of the Unvested Shares (rounded down to the nearest whole share) shall be scheduled to vest each month on the same day of the month as the Vesting Commencement Date , such that the Unvested Shares are vested in full as of October 16, 2027] /2[September 2, 2026 (the “Vesting Commencement Date”), 1/4th of the Unvested Shares (rounded down to the nearest whole share) shall be scheduled to vest each month on the same day of the month as the Vesting Commencement Date , such that the Unvested Shares are vested in full as of January 2, 2027]; provided in each case that Holder remains a Service Provider through each such date (the “Vesting Schedule”).
[Remainder of page intentionally left blank]
| 1 | NTD: Vesting schedule for Kathy Dong. |
| 2 | NTD: Vesting schedule for Graham Parry. |
Stock Restriction Agreement
[Holder]
Page 1
Additional Terms/Acknowledgements: The undersigned Holder acknowledges receipt of, and understands and agrees to, this Stock Restriction Agreement, including those terms and conditions set forth in Exhibit A attached to this Agreement, which are incorporated by reference.
| COMPANY: | ||||
| Electra Therapeutics, Inc. | ||||
| By: |
| |||
| Name: [ ] | ||||
| Title: [ ] | ||||
| Address: | 230 E Grand Avenue, Suite S-100 South San Francisco, California 94080 | |||
| HOLDER: | ||
| [ ] | ||
|
| ||
| (Signature) | ||
| Email: | ||
| Address: [ ] | ||
Stock Restriction Agreement
[Holder]
Signature Page
Exhibit A
TERMS AND CONDITIONS INCORPORATED INTO
STOCK RESTRICTION AGREEMENT
1. Consideration for Agreement. Holder has entered into this Agreement as a condition to receiving the Unvested Shares in connection with the distribution relating to Holder’s profits interests in Electra Therapeutics LLC (the “LLC Distribution”) and the Company’s initial public offering of its securities pursuant to a registration statement on Form S-1 (the “IPO”). Holder acknowledges and agrees that Holder would not have a right to receive the Unvested Shares in connection with the LLC Distribution if Holder did not enter into this Agreement. Holder also anticipates that the IPO will benefit Holder and acknowledges and agrees that Holder has received valid and sufficient consideration for entering into this Agreement.
2. Forfeiture of Unvested Shares. In the event Holder’s relationship with the Company (or a parent or subsidiary of the Company) terminates for any reason (including death or disability) or Holder resigns for any reason, such that after such termination Holder is no longer providing services to the Company (or a parent or subsidiary of the Company) as an employee, consultant or member of the Company’s Board of Directors (a “Service Provider”), then the Unvested Shares shall automatically and immediately be forfeited for no consideration and be retired and resume the status of authorized and unissued shares of the capital stock of the Company, without further action by Holder or the Company. Notwithstanding the foregoing, and for the avoidance of doubt, the foregoing will be subject to any severance plan or agreement to which Holder is a participant or a party.
3. Adjustments to Unvested Shares. If, from time to time, while the Unvested Shares remain subject to forfeiture due to being unvested there is any change affecting the Company’s outstanding common stock without the receipt of consideration by the Company (through merger, consolidation, reorganization, reincorporation, stock dividend, dividend in property other than cash, stock split, liquidating dividend, combination of shares, change in corporation structure or other transaction not involving the receipt of consideration by the Company), then any and all new, substituted or additional securities or other property to which Holder is entitled by reason of Holder’s ownership of Unvested Shares will be immediately subject to the Vesting Schedule applicable to the Unvested Shares and be included in the meaning of “Unvested Shares” for all purposes of this Agreement with the same force and effect as the Unvested Shares subject to this Agreement.
4. Escrow of Unvested Shares. As security for Holder’s faithful performance of the terms of this Agreement and to insure the availability for delivery of Holder’s Unvested Shares upon the forfeiture of any Unvested Shares as herein provided for, Holder agrees, concurrently with the execution of this Agreement, to deliver to and deposit with the Chief Financial Officer of the Company or the Chief Financial Officer’s designee, including the person or entity named in Joint Escrow Instructions of the Company and Holder attached to this Agreement as Exhibit C and incorporated by this reference (“Joint Escrow Instructions”), as Escrow Agent in this transaction (“Escrow Agent”), one stock assignment duly endorsed (with date and number of shares blank) in the form attached to this Agreement as Exhibit B, together with a certificate or certificates evidencing all Unvested Shares that are subject to forfeiture; said documents are to be held by the Escrow Agent and delivered by said Escrow Agent pursuant to the Joint Escrow Instructions, which instructions will also be delivered to the Escrow Agent concurrently with the execution of this Agreement. Holder acknowledges that the Escrow Agent is so appointed as the escrow holder with the foregoing authorities as a material inducement to make this Agreement and that said appointment is coupled with an interest and is accordingly irrevocable. Holder agrees that Escrow Agent is not liable to any party hereof (or to any other party). Escrow Agent may rely upon any letter, notice or other document executed by any signature purported to be genuine and may resign at any time. Holder agrees that if the Escrow Agent resigns as Escrow Agent for any or no reason, the Board of Directors of the Company has
Exhibit A to Stock Restriction Agreement
[Holder]
Page 1
the power to appoint a successor to serve as Escrow Agent pursuant to the terms of this Agreement. Holder agrees that if the Chief Financial Officer of the Company resigns as Chief Financial Officer, the successor Chief Financial Officer will serve as Escrow Agent pursuant to the terms of this Agreement.
5. Rights of Holder. Subject to the provisions of this Agreement, Holder will exercise all rights and privileges of a stockholder of the Company with respect to the Unvested Shares deposited in escrow. Holder will be deemed to be the holder for purposes of receiving any dividends that may be paid with respect to such Unvested Shares (but subject to Section 3) and for the purpose of exercising any voting rights relating to such Unvested Shares, even if some or all of such Unvested Shares remain subject to the Vesting Schedule. After any Unvested Shares have been vested and are no longer subject to forfeiture, Holder will not assign, hypothecate, donate, encumber or otherwise dispose of any interest in the Unvested Shares except in compliance with the provisions herein, the Company’s Bylaws and any other agreement to which the Holder is a party and applicable securities laws. Holder further acknowledges that Holder may be required to hold the Stock indefinitely. During the period of time during which the Holder holds the Stock, the value of the Stock may increase or decrease, and any risk associated with such Stock and such fluctuation in value will be borne by the Holder.
6. Limitations on Transfer. In addition to any other limitation on transfer created by applicable securities laws, Holder will not assign, hypothecate, donate, encumber or otherwise dispose of any interest in the Unvested Shares while the Unvested Shares are unvested and remain subject to vesting pursuant to the Vesting Schedule.
7. Corporate Transaction. In the event of (a) an Acquisition (as defined below); or (b) an Asset Transfer (as defined below) ((a) and (b) being collectively referred to in the Agreement as a “Corporate Transaction”), then to the extent this Agreement remains in effect with respect to any Unvested Shares, the rights of the Company under this Agreement will be assigned by the Company to any successor of the Company (or the successor’s parent) in connection with such Corporate Transaction. The forfeiture terms of this Agreement will apply to the new capital stock or other property received in exchange for the Unvested Shares in consummation of the Corporate Transaction, but only to the extent the Unvested Shares are at the time covered by such right. For the purposes of this Section 7: (i) “Acquisition” means (A) any consolidation or merger of the Company with or into any other corporation or other entity or person, or any other corporate reorganization; or (B) any transaction or series of related transactions to which the Company is a party in which in excess of 50% of the Company’s voting power is transferred; and (ii) “Asset Transfer” means a sale, lease, exclusive license or other disposition of all or substantially all of the assets of the Company.
8. Section 83(b) Election. The Holder shall consult with the Holder’s tax advisor to determine whether it would be appropriate for the Holder to make an election under Section 83(b) of the Internal Revenue Code of 1986, as amended (the “Code” and such election thereunder, an “83(b) Election”) with respect to the Unvested Shares. Any such 83(b) Election must be filed with the Internal Revenue Service within 30 days of this Agreement. Holder understands that failure to file such 83(b) Election in a timely manner may result in adverse U.S. federal income tax consequences for Holder. Holder further acknowledges and understands that it is Holder’s sole obligation and responsibility to timely file such 83(b) Election, and neither the Company nor the Company’s legal or financial advisors have any obligation or responsibility with respect to such filing. Holder acknowledges that the foregoing is only a summary of the effect of U.S. federal income taxation with respect to the Unvested Shares hereunder, and does not purport to be complete. Holder further acknowledges that the Company has directed Holder to seek independent advice regarding the applicable provisions of the Code, the income tax laws of any municipality, state or foreign country in which Holder may reside, and the tax consequences of Holder’s death, and that the Company is not making any representations or warranties regarding the tax consequences of this Agreement. Holder assumes all responsibility for filing such 83(b) Election and paying all taxes
Exhibit A to Stock Restriction Agreement
[Holder]
Page 2
resulting from such election or, if no 83(b) Election is made, taxes resulting from the lapse of the restrictions on the Unvested Shares. To the extent Holder files an 83(b) Election, Holder agrees to provide the Company with a copy of such filed 83(b) Election and proof of such filing.
9. Refusal to Transfer. The Company will not be required (i) to transfer on its books any Unvested Shares of the Company that have been transferred in violation of any of the provisions set forth in this Agreement or (ii) to treat as owner of such shares or to accord the right to vote as such owner or to pay dividends to any transferee to whom such shares have been so transferred.
10. No Employment Rights. This Agreement is not an employment or other service contract and nothing in this Agreement will affect in any manner whatsoever the right or power of the Company (or a parent or subsidiary of the Company) to terminate Holder’s employment or other service relationship for any reason at any time, with or without cause and with or without notice.
11. Restrictive Legends. All certificates representing the Unvested Shares will have endorsed thereon legends in substantially the following forms (in addition to any other legend which may be required by other agreements between the parties to this Agreement):
(a) “THE SHARES REPRESENTED BY THIS CERTIFICATE HAVE NOT BEEN REGISTERED UNDER THE SECURITIES ACT OF 1933 AS AMENDED. THEY MAY NOT BE SOLD, OFFERED FOR SALE, PLEDGED OR HYPOTHECATED IN THE ABSENCE OF AN EFFECTIVE REGISTRATION STATEMENT AS TO THE SECURITIES UNDER SAID ACT OR AN OPINION OF COUNSEL SATISFACTORY TO THE CORPORATION THAT SUCH REGISTRATION IS NOT REQUIRED.”
(b) “THE SHARES REPRESENTED BY THIS CERTIFICATE ARE SUBJECT TO A TRANSFER RESTRICTION, AS PROVIDED IN THE BYLAWS OF THE CORPORATION.”
(c) “THE SHARES REPRESENTED BY THIS CERTIFICATE ARE SUBJECT TO AN OPTION AND OTHER RESTRICTIONS AND CONDITIONS SET FORTH IN AN AGREEMENT BETWEEN THE CORPORATION AND THE REGISTERED HOLDER, OR SUCH HOLDER’S PREDECESSOR IN INTEREST, A COPY OF WHICH IS ON FILE AT THE PRINCIPAL OFFICE OF THE CORPORATION. ANY TRANSFER OR ATTEMPTED TRANSFER OF ANY SHARES SUBJECT TO SUCH OPTION IS VOID WITHOUT THE PRIOR EXPRESS WRITTEN CONSENT OF THE CORPORATION.”
(d) “THE SHARES REPRESENTED BY THIS CERTIFICATE ARE SUBJECT TO CERTAIN OBLIGATIONS WITH RESPECT TO FUTURE SECURITYHOLDERS’ AGREEMENTS SET FORTH IN AN AGREEMENT BETWEEN THE CORPORATION AND THE REGISTERED HOLDER, OR SUCH HOLDER’S PREDECESSOR IN INTEREST, A COPY OF WHICH IS ON FILE AT THE PRINCIPAL OFFICE OF THE CORPORATION.”
(e) Any legend required by applicable blue sky laws.
12. Miscellaneous.
(a) Notices. All notices required or permitted hereunder will be in writing and will be deemed effectively given: (i) upon personal delivery to the party to be notified; (ii) when sent by confirmed electronic mail if sent during normal business hours of the recipient, and if not sent during normal business hours of the recipient, then on the next business day (provided, however, that if the sender receives an automatically generated notification that such email was not delivered, such attempted email notice shall
Exhibit A to Stock Restriction Agreement
[Holder]
Page 3
be ineffective and deemed to not have been given); (iii) five calendar days after having been sent by registered or certified mail, return receipt requested, postage prepaid; or (iv) one business day after deposit with a nationally recognized overnight courier, specifying next day delivery, with written verification of receipt. All communications will be sent to the other party to this Agreement at such party’s address set forth on the signature page hereof, or at such other address as such party may designate by 10 days’ advance written notice to the other party hereto.
(b) Successors and Assigns. This Agreement will inure to the benefit of the successors and assigns of the Company and, subject to the restrictions on transfer herein set forth, be binding upon Holder, Holder’s successors, and assigns. The Company’s rights hereunder will be assignable by the Company at any time or from time to time, in whole or in part.
(c) Attorneys’ Fees. The prevailing party in any suit or action hereunder will be entitled to recover from the losing party all costs incurred by it in enforcing the performance of, or protecting its rights under, any part of this Agreement, including reasonable costs of investigation and attorneys’ fees.
(d) Governing Law; Venue. This Agreement will be governed by and construed in accordance with the laws of the State of Delaware. The parties agree that any action brought by either party to interpret or enforce any provision of this Agreement will be brought in, and each party agrees to, and does hereby, submit to the jurisdiction and venue of, the appropriate state or federal court for the district encompassing the Company’s principal place of business.
(e) Further Execution. The parties agree to take all such further actions as may reasonably be necessary to carry out and consummate this Agreement as soon as practicable, and to take whatever steps may be necessary to obtain any governmental approval in connection with or otherwise qualify the issuance of the securities that are the subject of this Agreement. Holder agrees that, at the request of the Company at any time, Holder will execute and deliver any applicable securityholders’ agreement, investor rights agreement, voting agreement, drag along agreement, right of first refusal and co-sale agreement or similar agreement (or a joinder to any existing agreement) that the Company and/or the holders of its securities may enter into or that otherwise that may be in effect from time to time (and which may contain, among other provisions, additional restrictions on transfer and/or rights of repurchase in favor of the Company).
(f) Entire Agreement; Amendment. This Agreement constitutes the entire agreement between the parties with respect to the subject matter hereof and supersedes and merges all prior agreements or understandings, whether written or oral, with respect to the subject matter hereof. This Agreement may not be amended, modified or revoked, in whole or in part, except by an agreement in writing signed by each of the parties hereto.
(g) Severability. If one or more provisions of this Agreement are held to be unenforceable under applicable law, the parties agree to renegotiate such provision in good faith. In the event that the parties cannot reach a mutually agreeable and enforceable replacement for such provision, then (i) such provision will be excluded from this Agreement, (ii) the balance of the Agreement will be interpreted as if such provision were so excluded and (iii) the balance of the Agreement will be enforceable in accordance with its terms.
(h) Counterparts. This Agreement (including any schedules and/or exhibits hereto or thereto) may be executed in two or more counterparts, each of which will be deemed an original, but all of which together will constitute one and the same instrument. Counterparts may be delivered via facsimile, electronic mail (including pdf or any electronic signature complying with the U.S. federal ESIGN Act of 2000, Uniform Electronic Transactions Act or other applicable law) or other transmission method and any
Exhibit A to Stock Restriction Agreement
[Holder]
Page 4
counterpart so delivered will be deemed to have been duly and validly delivered and be valid and effective for all purposes.
(i) Tax Matters. Holder expressly acknowledges and agrees that (i) neither the Company, nor any of its affiliates or agents, has made any representation to Holder with respect to the tax or other financial treatment of the transactions contemplated by this Agreement, (ii) Holder has reviewed with its own tax advisors the tax consequences of the transactions contemplated by this Agreement, and (iii) Holder is relying solely on such advisors and not on any statements or representations of the Company or any of its affiliates or agents for the tax consequences to Holder that may result from the transactions contemplated by this Agreement. Holder shall be solely responsible for the payment of any and all income, capital gain, payroll, employment, transfer, and other taxes, filing and recording fees, and similar charges incurred by Holder, and any income or withholding taxes incurred, with respect to payments (including payments of property) made to Holder pursuant to the transactions contemplated by this Agreement (regardless of if such taxes would normally be paid or withheld by an employer or other withholding agent), and shall indemnify and hold harmless the Company from and against any such taxes, fees, and charges (including any interest and penalties, but excluding any employer payroll taxes). Holder shall provide any tax forms, including Internal Revenue Service Form W-9, or any similar form or information, as may be reasonably necessary to reduce or eliminate any tax withholding, and Holder expressly acknowledges that the Company may deduct and withhold from payments made to Holder pursuant to the transactions contemplated by this Agreement (or otherwise) such amounts as may be required to be deducted or withheld under applicable law, including if Holder fails to provide such tax forms. To the extent amounts are so deducted or withheld, such amounts shall be treated for all purposes of this Agreement as having been paid to Holder.
[End of Exhibit A to Stock Restriction Agreement]
Exhibit A to Stock Restriction Agreement
[Holder]
Page 5
Exhibit B
STOCK ASSIGNMENT SEPARATE FROM CERTIFICATE
Effective upon forfeiture under the certain Stock Restriction Agreement, dated September __, 2026, by and between the undersigned and Electra Therapeutics, Inc, a Delaware corporation (the “Company”)(the “Agreement”) and for no consideration, the undersigned assigns and transfers to the Company pursuant to the Agreement ________________shares of Series A Preferred Stock of the Company (or such security into which such shares may convert) standing in the undersigned’s name on the books of the Company in book entry form and does irrevocably constitute and appoint both the Company’s Secretary and the Company’s attorney, or either of them, to transfer said stock on the books of the Company with full power of substitution in the premises. This Assignment may be used only in accordance with and subject to the terms and conditions of the Agreement, solely to evidence and effectuate the forfeiture, cancellation and retirement of Unvested Shares under the Agreement, and only to the extent that such shares constitute Unvested Shares subject to forfeiture under the Agreement.
| Dated: |
|
[HOLDER] | ||||||
| (Leave blank) | ||||||||
|
| ||||||||
| (Signature) |
Instruction: Please do not fill in any blanks other than the signature line. Do not fill in the date line. The purpose of this Assignment is to enable the Company to evidence and effectuate the forfeiture, cancellation and retirement of Unvested Shares under the Agreement without requiring additional signatures on the part of the Holder.
[Holder]
Page 6
ELECTRA THERAPEUTICS, INC.
JOINT ESCROW INSTRUCTIONS
Chief Financial Officer
Electra Therapeutics, Inc.
230 E Grand Avenue, Suite S-100
South San Francisco, CA 94080
As Escrow Agent for both Electra Therapeutics, Inc., a Delaware Company (“Company”) and [_______] (“Holder”), you are hereby authorized and directed to hold the documents delivered to you pursuant to the terms of that certain Stock Restriction Agreement dated as of September __, 2026 (“Agreement”), to which a copy of these Joint Escrow Instructions is attached as an Exhibit, in accordance with the following instructions:
1. In the event any Unvested Shares are subject to forfeiture as set forth in the Agreement, the Company or its assignee will give to Holder and you a written notice specifying the number of shares of stock to be forfeited and delivered to the Company for cancellation. Holder and the Company hereby irrevocably authorize and direct you to close the transaction contemplated by such notice in accordance with the terms of said notice.
2. At the closing, you are directed (a) to date the stock assignments necessary for the transfer in question, (b) to fill in the number of shares being transferred, and (c) to deliver the same, together with the certificate evidencing the shares of stock to be transferred, to the Company.
3. Holder irrevocably authorizes the Company to deposit with you any certificates evidencing shares of stock to be held by you hereunder and any additions and substitutions to said shares as specified in the Agreement. Holder does hereby irrevocably constitute and appoint you as Holder’s attorney-in-fact and agent for the term of this escrow to execute with respect to such securities all documents necessary or appropriate to make such securities negotiable and complete any transaction herein contemplated, including but not limited to any appropriate filing with state or government officials or bank officials. Subject to the provisions of this Section 3, Holder will exercise all rights and privileges of a stockholder of the Company while the stock is held by you.
4. This escrow will terminate upon the vesting in full of the Unvested Shares or the forfeiture of any Unvested Shares pursuant to the Agreement, whichever occurs first.
5. If at the time of termination of this escrow under Section 4 herein you should have in your possession any documents, securities, or other property belonging to Holder, you will deliver all of the same to Holder and will be discharged of all further obligations hereunder; provided, however, that if at the time of termination of this escrow you are advised by the Company that any property subject to this escrow is the subject of a pledge or other security agreement, you will deliver all such property to the pledgeholder or other person designated by the Company.
6. Except as otherwise provided in these Joint Escrow Instructions, your duties hereunder may be altered, amended, modified or revoked only by a writing signed by all of the parties hereto.
7. You are obligated only for the performance of such duties as are specifically set forth herein and may rely and will be protected in relying or refraining from acting on any instrument reasonably believed by you to be genuine and to have been signed or presented by the proper party or parties. You
Joint Escrow Instructions
[ ]Page 7
will not be personally liable for any act you may do or omit to do hereunder as Escrow Agent or as attorney-in-fact for Holder while acting in good faith and in the exercise of your own good judgment, and any act done or omitted by you pursuant to the advice of your own attorneys will be conclusive evidence of such good faith.
8. You are hereby expressly authorized to disregard any and all warnings given by any of the parties hereto or by any other person or entity, excepting only orders or process of courts of law, and are hereby expressly authorized to comply with and obey orders, judgments or decrees of any court. In case you obey or comply with any such order, judgment or decree of any court, you will not be liable to any of the parties hereto or to any other person, firm or corporation by reason of such compliance, notwithstanding any such order, judgment or decree being subsequently reversed, modified, annulled, set aside, vacated or found to have been entered without jurisdiction.
9. You will not be liable in any respect on account of the identity, authorities or rights of the parties executing or delivering or purporting to execute or deliver these Joint Escrow Instructions documents or papers deposited or called for hereunder.
10. You will not be liable for the outlawing of any rights under any statute of limitations with respect to these Joint Escrow Instructions or any documents deposited with you.
11. Your responsibilities as Escrow Agent hereunder will terminate if you cease to be Chief Financial Officer of the Company or if you resign by written notice to the Company. In the event of any such termination, the Chief Financial Officer of the Company will automatically become the successor Escrow Agent unless the Company appoints another successor Escrow Agent, and Holder hereby confirms the appointment of such successor as Holder’s attorney-in-fact and agent to the full extent of your appointment.
12. If you reasonably require other or further instruments in connection with these Joint Escrow Instructions or obligations in respect hereto, the necessary parties hereto will join in furnishing such instruments.
13. It is understood and agreed that should any dispute arise with respect to the delivery and/or ownership or right of possession of the securities held by you hereunder, you are authorized and directed to retain in your possession without liability to anyone all or any part of said securities until such dispute has been settled either by mutual written agreement of the parties concerned or by a final order, decree or judgment of a court of competent jurisdiction after the time for appeal has expired and no appeal has been perfected, but you will be under no duty whatsoever to institute or defend any such proceedings.
14. All notices required or permitted hereunder will be in writing and will be deemed effectively given: (a) upon personal delivery to the party to be notified, (b) when sent by email if sent during normal business hours of the recipient, and if not sent during normal business hours of the recipient, then on the next business day (provided, however, that if the sender receives an automatically generated notification that such email was not delivered, such attempted email notice shall be ineffective and deemed to not have been given), (c) five calendar days after having been sent by registered or certified mail, return receipt requested, postage prepaid, or (d) one business day after deposit with a nationally recognized overnight courier, specifying next day delivery, with written verification of receipt. All communications will be sent to the other party hereto at such party’s address set forth on the signature page to these Joint Escrow Instructions, or at such other address as such party may designate by 10 days’ advance written notice to the other party hereto.
Joint Escrow Instructions
[ ]Page 8
15. By signing these Joint Escrow Instructions, you become a party hereto only for the purpose of said Joint Escrow Instructions; you do not become a party to the Agreement.
16. You are entitled to employ such legal counsel and other experts (including, without limitation, the firm of Cooley LLP) as you may deem necessary properly to advise you in connection with your obligations hereunder. You may rely upon the advice of such counsel, and you may pay such counsel reasonable compensation therefor.
17. This instrument is binding upon and inures to the benefit of the parties hereto and their respective successors and permitted assigns. It is understood and agreed that references to “you” and “your” herein refer to the original Escrow Agents and to any and all successor Escrow Agents. It is understood and agreed that the Company may at any time or from time to time assign its rights under the Agreement and these Joint Escrow Instructions in whole or in part.
18. These Joint Escrow Instructions will be governed by and interpreted and determined in accordance with the laws of the State of Delaware, as such laws are applied by Delaware courts to contracts made and to be performed entirely in Delaware by residents of that state. The parties hereby expressly consent to the personal jurisdiction of the state and federal courts located in the county in which the Company has its principal offices for any lawsuit arising from or related to this Agreement.
[Remainder of page intentionally left blank]
Joint Escrow Instructions
[ ]Page 9
The undersigned have executed these Joint Escrow Instructions as of the date set forth above.
| HOLDER: | ||
| [ ] | ||
|
| ||
| (Signature) | ||
| Address: [ ] | ||
| Email: | ||
| COMPANY: | ||||
| Electra Therapeutics, Inc. | ||||
| By: |
| |||
| Name: [ ] | ||||
| Title: [ ] | ||||
ESCROW AGENT:
| By: |
| |||
| Name: [ ] | ||||
| Title: Chief Financial Officer | ||||
Joint Escrow Instructions
[ ]Page 10
Exhibit 10.15
OPTION CANCELLATION AGREEMENT
This OPTION CANCELLATION AGREEMENT (this “Agreement”) is made and entered into on September 11, 2026, by and between Electra Therapeutics, Inc. (the “Company”) and Quehuong (Kathy) Dong, Pharm.D., M.B.A. (“Optionee”).
RECITALS
WHEREAS, on February 13, 2026, the Company granted to Optionee options to purchase 342,048 shares of Company common stock (as adjusted for any stock split, the “Option”) under the terms of the Company’s 2022 Stock Plan;
WHEREAS, the Option remains outstanding as of the date of this Agreement; and
WHEREAS, in connection with the Company’s underwritten initial public offering (the “IPO”), the Company and Optionee desire to cancel the entire amount of the Option contingent upon and concurrent with the Company’s grant of a stock option to acquire 927,155 shares of the Company’s common stock (the “IPO Grant”), a portion of which is being granted to the Optionee as consideration for entering into this Agreement (the date of IPO Grant, the “Effective Date,” and the time at which such IPO Grant is made, the “Effective Time”).
AGREEMENT
NOW, THEREFORE, in consideration of the mutual promises and covenants and conditions herein and for good and valuable consideration, receipt and sufficiency of which are hereby acknowledged, the parties hereto mutually agree as follows:
1. Cancellation of Option. The Company and Optionee agree that it is in the best interests of the Company and Optionee, as a holder of equity interests in the Company, to cancel the Option. As a result of the foregoing, the Company and the Optionee hereby agree that the Option is hereby terminated and cancelled effective as of the Effective Time.
2. No Further Rights. Optionee (i) acknowledges and agrees that except as otherwise set forth in this Agreement, Optionee is not entitled to any value, nor expects any value, in consideration for the cancellation of the Option, (ii) waives any and all rights Optionee may have had, now has or may ever have with respect to the Option, including any economic value of the Option, (iii) agrees and acknowledges that the Option will be of no further force or effect as of the Effective Time, (iv) hereby forfeits the Option in full, and (v) irrevocably and unconditionally releases the Company from any and all charges, complaints, claims, liabilities, obligations, promises, agreements, controversies, damages or causes of action, suits, rights, demands, costs, losses, debts and expenses (including attorneys’ fees and costs incurred) of any nature whatsoever, known or unknown, suspected or unsuspected, that Optionee may have had in the past, may now have or may have relating to the Option.
3. Release and Waiver of Claims. Effective as of the Effective Date, Optionee releases the Company and its officers, directors, employees, shareholders, parents, subsidiaries, affiliates, successors, assigns, agents, and attorneys, from any and all claims, liabilities, demands, causes of action, attorneys’ fees, damages, or obligations of every kind and nature, whether they are now known or unknown, arising in connection with the Option on or prior to the Effective Date.
1
4. Further Assurances. Optionee agrees to execute and/or cause to be delivered to the Company such instruments and other documents, and shall take such other actions, as the Company may reasonably request for the purpose of carrying out or evidencing the provisions of this Agreement.
5. Legal and Tax Advice. The Company makes no representation as to the tax consequences to Optionee of the transactions contemplated by this Agreement, which shall be the sole responsibility of Optionee. Optionee acknowledges and agrees that Optionee has had the opportunity to consult with Optionee’s personal legal and tax advisors in connection with this Agreement and that Optionee is not relying upon the Company or its counsel for any legal or tax advice.
6. Entire Agreement and Amendments. This Agreement constitutes the entire agreement of the parties with respect to the subject matter hereof and may not be modified, amended or terminated except by an agreement signed by the parties hereto. This Agreement shall supersede all prior agreements, arrangements and understandings, whether written or oral, between the parties regarding the subject matter of this Agreement.
7. Governing Law; Successor and Assigns. This Agreement shall be governed in all respects by the laws of the State of Delaware without regard to its conflicts of laws principles and shall be binding upon the heirs, personal representatives, executors, administrators, successors and assigns of the parties.
8. Counterparts. This Agreement may be executed and delivered in counterparts and by any electronic signature complying with the U.S. federal ESIGN Act of 2000, each of which will be deemed an original and all of which together will constitute one and the same instrument.
[Signature Page Follows]
2
IN WITNESS WHEREOF, this Agreement has been executed by the parties hereto as of the date first above written.
| ELECTRA THERAPEUTICS, INC. | ||
| By: | /s/ Nancy Stagliano, Ph.D. | |
| Name: | Nancy Stagliano, Ph.D. | |
| Title: | Chairperson of the Board | |
| OPTIONEE |
| /s/ Quehuong (Kathy) Dong, Pharm.D., M.B.A. |
| Name: Quehuong (Kathy) Dong, Pharm.D., M.B.A. |
3
Exhibit 23.1
Consent of Independent Registered Public Accounting Firm
We consent to the reference to our firm under the caption “Experts” and to the use of our report dated June 26, 2026 (except for the seventh paragraph of Note 1, as to which the date is September 14, 2026) in the Amendment No. 1 to the Registration Statement (Form S-1 No. 333-298617) and related Prospectus of Electra Therapeutics, Inc. for the registration of shares of its common stock.
/s/ Ernst & Young LLP
San Mateo, California
September 14, 2026
| Calculation of Filing Fee Tables | |||
| | |||
| | |||
| Table 1: Newly Registered and Carry Forward Securities |
|---|
| Security Type |
Security Class Title |
Fee Calculation or Carry Forward Rule |
Amount Registered |
Proposed Maximum Offering Price Per Unit |
Maximum Aggregate Offering Price |
Fee Rate |
Amount of Registration Fee |
Carry Forward Form Type |
Carry Forward File Number |
Carry Forward Initial Effective Date |
Filing Fee Previously Paid in Connection with Unsold Securities to be Carried Forward | ||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Newly Registered Securities | |||||||||||||
| |
1 | |
|
|
|
$ |
$ |
|
$ |
||||
| |
2 | |
|
|
|
$ |
$ |
$ |
|||||
| Carry Forward Securities | |||||||||||||
| Carry Forward Securities | |||||||||||||
| Total Offering Amounts: |
$ |
$ |
|||||||||||
| Total Fees Previously Paid: |
$ |
||||||||||||
| Total Fee Offsets: |
$ |
||||||||||||
| Net Fee Due: |
$ |
||||||||||||
| Offering Note |
| 1 |
| ||||||
| | |||||||
| 2 |
| ||||||
| | |||||||
| Table 2: Fee Offset Claims and Sources |
|---|
| Registrant or Filer Name | Form or Filing Type | File Number | Initial Filing Date | Filing Date | Fee Offset Claimed | Security Type Associated with Fee Offset Claimed | Security Title Associated with Fee Offset Claimed | Unsold Securities Associated with Fee Offset Claimed | Unsold Aggregate Offering Amount Associated with Fee Offset Claimed | Fee Paid with Fee Offset Source | |||
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| Rules 457(b) and 0-11(a)(2) | |||||||||||||
| Fee Offset Claims | |||||||||||||
| Fee Offset Sources | |||||||||||||
| Rule 457(p) | |||||||||||||
| Fee Offset Claims | |||||||||||||
| Fee Offset Sources | |||||||||||||
| Table 3: Combined Prospectuses |
|---|
| Security Type |
Security Class Title |
Amount of Securities Previously Registered |
Maximum Aggregate Offering Price of Securities Previously Registered |
Form Type |
File Number |
Initial Effective Date | |
|---|---|---|---|---|---|---|---|